WO2020122182A1 - 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 - Google Patents
分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 Download PDFInfo
- Publication number
- WO2020122182A1 WO2020122182A1 PCT/JP2019/048720 JP2019048720W WO2020122182A1 WO 2020122182 A1 WO2020122182 A1 WO 2020122182A1 JP 2019048720 W JP2019048720 W JP 2019048720W WO 2020122182 A1 WO2020122182 A1 WO 2020122182A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- group
- compound
- formula
- hydrogen
- amino acid
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 CC(C)C[C@@](C(N[C@@](CC(N([C@@]1C)N2[C@@](Cc3ccccc3)C(N[C@@](C)C(N3CCCCC3)=O)=O)=*=C1N[C@@](Cc1ccccc1)C(N(C)[C@@](CC(C)C)C(N[C@@](Cc1c[s]cn1)C(N(C)[C@@](Cc1ccccc1)C(NCC(N(C)[C@@](CC(C)C)C(N(C)[C@]1CC(C)C)=O)=O)=O)=O)=O)=O)C2=O)=O)NC1=O Chemical compound CC(C)C[C@@](C(N[C@@](CC(N([C@@]1C)N2[C@@](Cc3ccccc3)C(N[C@@](C)C(N3CCCCC3)=O)=O)=*=C1N[C@@](Cc1ccccc1)C(N(C)[C@@](CC(C)C)C(N[C@@](Cc1c[s]cn1)C(N(C)[C@@](Cc1ccccc1)C(NCC(N(C)[C@@](CC(C)C)C(N(C)[C@]1CC(C)C)=O)=O)=O)=O)=O)=O)C2=O)=O)NC1=O 0.000 description 2
- HPUIBRYCCDZJIC-AKEBHFNQSA-N CC(C)C[C@@H](C(N(C)CC(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](C(C)C)C(N[C@@H](CCc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N(C)[C@@H](C(C)C)C(NC(C(N(C)[C@H]1Cc2ccccc2)=O)=C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(N(C)CC(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](C(C)C)C(N[C@@H](CCc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N(C)[C@@H](C(C)C)C(NC(C(N(C)[C@H]1Cc2ccccc2)=O)=C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O HPUIBRYCCDZJIC-AKEBHFNQSA-N 0.000 description 1
- AUELSOQYRISSCI-QKWQJRFLSA-N CC(C)C[C@@H](C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(N(CCC1)[C@@H]1C(N(C)[C@H]1Cc2ccccc2)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O Chemical compound CC(C)C[C@@H](C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(N(CCC1)[C@@H]1C(N(C)[C@H]1Cc2ccccc2)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O AUELSOQYRISSCI-QKWQJRFLSA-N 0.000 description 1
- JYUCCSPOAZJQNB-DBPNMAKSSA-N CC(C)C[C@@H](C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CCc1ccc(C(NC)=O)c(OC)c1)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](C(C)C)C(N(C)[C@H]1Cc2ccccc2)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O Chemical compound CC(C)C[C@@H](C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CCc1ccc(C(NC)=O)c(OC)c1)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](C(C)C)C(N(C)[C@H]1Cc2ccccc2)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O JYUCCSPOAZJQNB-DBPNMAKSSA-N 0.000 description 1
- AQJXUWRLJSXPJP-VJCSGHBVSA-N CC(C)C[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O Chemical compound CC(C)C[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O AQJXUWRLJSXPJP-VJCSGHBVSA-N 0.000 description 1
- NQLRWLWIAKNWSN-KDTWXVJPSA-N CC(C)C[C@@H](C(NC(COCC(C)(C)CO)C(N(C)C(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N/C(/C(N(C)[C@@H](Cc1ccccc1)C(N(C)/C(/C(NC([C@@H](C)O)C(N(C)CC(N(C)C(CC(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=C\C(C)C)=O)=O)=C/C(C)C)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(NC(COCC(C)(C)CO)C(N(C)C(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N/C(/C(N(C)[C@@H](Cc1ccccc1)C(N(C)/C(/C(NC([C@@H](C)O)C(N(C)CC(N(C)C(CC(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=C\C(C)C)=O)=O)=C/C(C)C)=O)=O)=O)=O)N(C)C1=O NQLRWLWIAKNWSN-KDTWXVJPSA-N 0.000 description 1
- BJCKCMMCZJNJRY-CLRMZFCCSA-N CC(C)C[C@@H](C(N[C@@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCC1)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(N[C@@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCC1)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O BJCKCMMCZJNJRY-CLRMZFCCSA-N 0.000 description 1
- DFAQTPPZWBGHPY-TZDMAYCNSA-N CC(C)C[C@@H](C(N[C@@H](CCc(cc1)cc(OC)c1C(NC)=O)C(N(C)C(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCC1)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(N[C@@H](CCc(cc1)cc(OC)c1C(NC)=O)C(N(C)C(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCC1)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O DFAQTPPZWBGHPY-TZDMAYCNSA-N 0.000 description 1
- ZXDCQXVNXUFIGS-VRYVOHGFSA-N CC(C)C[C@@H](C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@@H](Cc1ccccc1)C(NC(COc1cccc(Cl)c1)C(N(C)[C@@H](C(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)N(C)C2=[O]C2CN(C)C1=O Chemical compound CC(C)C[C@@H](C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@@H](Cc1ccccc1)C(NC(COc1cccc(Cl)c1)C(N(C)[C@@H](C(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)N(C)C2=[O]C2CN(C)C1=O ZXDCQXVNXUFIGS-VRYVOHGFSA-N 0.000 description 1
- UPLJMTTZDNPYJM-NRXHODDDSA-N CC(C)C[C@@H](C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CCc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@H]1CCc2ccccc2)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CCc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@H]1CCc2ccccc2)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O UPLJMTTZDNPYJM-NRXHODDDSA-N 0.000 description 1
- VIIFORDSOUKRPR-WXLSUELNSA-N CC(C)C[C@@H](C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@H]1CCc2ccccc2)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC(C)C[C@@H](C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@H]1CCc2ccccc2)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O VIIFORDSOUKRPR-WXLSUELNSA-N 0.000 description 1
- ZCDYTNZJBGSKFI-UHFFFAOYSA-N CC(c1c(C)cccc1)N Chemical compound CC(c1c(C)cccc1)N ZCDYTNZJBGSKFI-UHFFFAOYSA-N 0.000 description 1
- CBYBHLAAQBWFSN-OWCCKUAASA-N CCCCC(CC)C[C@@H](C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@H]1CC(C)C)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)NC1=O Chemical compound CCCCC(CC)C[C@@H](C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@H]1CC(C)C)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)NC1=O CBYBHLAAQBWFSN-OWCCKUAASA-N 0.000 description 1
- UDOJNAFSQOHLBO-OFESFFRBSA-N CCCCN(CC(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COCC[C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COCC[C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O UDOJNAFSQOHLBO-OFESFFRBSA-N 0.000 description 1
- HHHMQKGBQLMPPZ-CKZABATQSA-N CCCCN(CC(N(C)[C@@H](CC(C)C)C(NC[C@@H](COCCO)C(N(C)/C(/C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](CC(C)C)C(N(C)[C@@H](CC(C)C)CN[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=C/c2ccccc2)=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](CC(C)C)C(NC[C@@H](COCCO)C(N(C)/C(/C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N[C@H](CC(C)C)C(N(C)[C@@H](CC(C)C)CN[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=C/c2ccccc2)=O)=O)=O)C1=O HHHMQKGBQLMPPZ-CKZABATQSA-N 0.000 description 1
- QXJMTNBZDWZANA-VWSFQUFSSA-N CCCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N(C)/C(/C(N[C@@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=C\c2ccccc2)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N(C)/C(/C(N[C@@H](C(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=C\c2ccccc2)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)C1=O QXJMTNBZDWZANA-VWSFQUFSSA-N 0.000 description 1
- LAKMXHCQDUOTDG-TWJDDVTBSA-N CCCCN(CC(N(C)[C@@H](Cc1cccc(Cl)c1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](COCC[C@H](C)O)C(N(C)[C@@H](CC(C)C)C(NC(CC(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](Cc1cccc(Cl)c1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](COCC[C@H](C)O)C(N(C)[C@@H](CC(C)C)C(NC(CC(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O LAKMXHCQDUOTDG-TWJDDVTBSA-N 0.000 description 1
- NIBJFGOECABCIC-QQCOAIBHSA-N CCCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CC(N(C)[C@@H](Cc1cccc(Cl)c1)C(N[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](C(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CC(N(C)[C@@H](Cc1cccc(Cl)c1)C(N[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O NIBJFGOECABCIC-QQCOAIBHSA-N 0.000 description 1
- DXNMIXCKAYWEBI-LHFWSPFISA-N CCCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@](CC(C)C)(CN[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)C=O)=O)=O)C1=O Chemical compound CCCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@](CC(C)C)(CN[C@@H](COCC(C)(C)CO)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)C=O)=O)=O)C1=O DXNMIXCKAYWEBI-LHFWSPFISA-N 0.000 description 1
- ZQLMNBDEJVKTIY-BPBYABOKSA-N CCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(NC(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(NC[C@@H](COCCO)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(NC(CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(NC[C@@H](COCCO)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O ZQLMNBDEJVKTIY-BPBYABOKSA-N 0.000 description 1
- SVFSVBIZLIBYJP-KWDANQMSSA-N CCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCN(CC(N(C)[C@@H](CC(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(N[C@H](C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](C(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@H]1[C@@H](C)O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)C1=O SVFSVBIZLIBYJP-KWDANQMSSA-N 0.000 description 1
- QEQYKFURPASBDQ-KAXMKOBWSA-N CCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)CC)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)C1=O Chemical compound CCCN(CC(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)CC)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](COC(C)(C)C)C(N(C)[C@@H](C(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)C1=O QEQYKFURPASBDQ-KAXMKOBWSA-N 0.000 description 1
- RTZYEAOMDPFIBT-FIQDSTCUSA-N CC[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@](CN[C@@H](CCCC(C(F)(F)F)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)(C(C)C)C=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@](CN[C@@H](CCCC(C(F)(F)F)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)(C(C)C)C=O)=O)=O)=O)=O)N(C)C1=O RTZYEAOMDPFIBT-FIQDSTCUSA-N 0.000 description 1
- LOJASVLPVQXALK-ZHGDGUHFSA-N CC[C@@H](C(N[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(NC(COCC(C)(C)CO)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC[C@@H](C(N[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(NC(COCC(C)(C)CO)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](C1)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)=O)N(C)C1=O LOJASVLPVQXALK-ZHGDGUHFSA-N 0.000 description 1
- VGLZFKCRXBITOG-CTKORFLVSA-N CC[C@H](C)[C@@H](C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCCC(N(C)[C@@H](CC)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@H]1CC(C)C)=O)=O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)NC1=O Chemical compound CC[C@H](C)[C@@H](C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCCC(N(C)[C@@H](CC)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](COC(C)(C)C)C(N(C)[C@H]1CC(C)C)=O)=O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)NC1=O VGLZFKCRXBITOG-CTKORFLVSA-N 0.000 description 1
- OSPWUODJPVGFBY-UGPVPSPFSA-N CC[C@H](C)[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CCc(cc1)cc(OC)c1C(NC)=O)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCCC(N(C)[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O Chemical compound CC[C@H](C)[C@@H](C(N(C)[C@@H](Cc1ccccc1)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CCc(cc1)cc(OC)c1C(NC)=O)C(N(C)[C@@H](CC(C)C)C(N[C@@H]([C@@H](C)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)CCCC(N(C)[C@H]1CC(C)C)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)=O)=O)=O)=O)NC1=O OSPWUODJPVGFBY-UGPVPSPFSA-N 0.000 description 1
- CCTPMTIURKHUDG-MULLBPGRSA-N CC[C@H](C)[C@@H](C(N[C@@H](C(C)C)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@H]1Cc2ccccc2)=O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)N(C)C1=O Chemical compound CC[C@H](C)[C@@H](C(N[C@@H](C(C)C)C(N(C)CC(N(C)[C@@H](CC(C)C)C(N[C@@H](CC(N(C)[C@@H](C)C(N[C@@H](CCCC(C(F)(F)F)O)C(N(C)[C@@H](CC(C)C)C(N(C)[C@@H](Cc1ccccc1)C(N[C@@H](CC(C)C)C(N(C)[C@H]1Cc2ccccc2)=O)=O)=O)=O)=O)=O)C(N2CCCCC2)=O)=O)=O)=O)=O)N(C)C1=O CCTPMTIURKHUDG-MULLBPGRSA-N 0.000 description 1
- VNMHEZQICDRVEM-WOJGMQOQSA-N CN(/C(/C(O)=O)=C/OCC(C(F)(F)F)O)C(OCC1C(C=CCC2)=C2c2ccccc12)=O Chemical compound CN(/C(/C(O)=O)=C/OCC(C(F)(F)F)O)C(OCC1C(C=CCC2)=C2c2ccccc12)=O VNMHEZQICDRVEM-WOJGMQOQSA-N 0.000 description 1
- CJOISDUTEOTEHX-MBMZGMDYSA-N CN(C(c1ccc(C[C@@H](C(O)=O)NC(OCC2c3ccccc3-c3c2cccc3)=O)cc1)=O)OC1OCCCC1 Chemical compound CN(C(c1ccc(C[C@@H](C(O)=O)NC(OCC2c3ccccc3-c3c2cccc3)=O)cc1)=O)OC1OCCCC1 CJOISDUTEOTEHX-MBMZGMDYSA-N 0.000 description 1
- HTDWMNDUFNXBIA-FQEVSTJZSA-N CN([C@@H](CC(OCC=C)=O)C(O)=O)C(OCC(c1ccccc1-1)c2c-1cccc2)=O Chemical compound CN([C@@H](CC(OCC=C)=O)C(O)=O)C(OCC(c1ccccc1-1)c2c-1cccc2)=O HTDWMNDUFNXBIA-FQEVSTJZSA-N 0.000 description 1
- MWJPOLRMZHFFLL-SFHVURJKSA-N CNC(CC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2ccccc12)=O)=O Chemical compound CNC(CC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2ccccc12)=O)=O MWJPOLRMZHFFLL-SFHVURJKSA-N 0.000 description 1
- IOHSXVPCMIYNCL-SFHVURJKSA-N CNC(COC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2c1cccc2)=O)=O Chemical compound CNC(COC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2c1cccc2)=O)=O IOHSXVPCMIYNCL-SFHVURJKSA-N 0.000 description 1
- KLTLIGBOCPHNGZ-LBPRGKRZSA-N COC([C@H](CCCC=O)N(C(c1c2cccc1)=O)C2=O)=O Chemical compound COC([C@H](CCCC=O)N(C(c1c2cccc1)=O)C2=O)=O KLTLIGBOCPHNGZ-LBPRGKRZSA-N 0.000 description 1
- NJDSRMVYAFOBIZ-FTNKSUMCSA-N C[C@@H]1OC(c2ccccc2)OCC1 Chemical compound C[C@@H]1OC(c2ccccc2)OCC1 NJDSRMVYAFOBIZ-FTNKSUMCSA-N 0.000 description 1
- SEINCOLPXCLPDL-SNVBAGLBSA-N C[C@H](CCO)OCc1ccccc1 Chemical compound C[C@H](CCO)OCc1ccccc1 SEINCOLPXCLPDL-SNVBAGLBSA-N 0.000 description 1
- XPDFLMWIBWUCRC-XLIONFOSSA-N C[C@H](COC[C@@H](C(O)=O)N(C)C(OCc1ccccc1)=O)OCc1ccccc1 Chemical compound C[C@H](COC[C@@H](C(O)=O)N(C)C(OCc1ccccc1)=O)OCc1ccccc1 XPDFLMWIBWUCRC-XLIONFOSSA-N 0.000 description 1
- LELUAYLNMNZFCY-UHNVWZDZSA-N C[C@H](COC[C@@H](C(O)=O)N)O Chemical compound C[C@H](COC[C@@H](C(O)=O)N)O LELUAYLNMNZFCY-UHNVWZDZSA-N 0.000 description 1
- SXLQXHDUCLWZIE-YJYMSZOUSA-N C[C@H](COC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2ccccc12)=O)O Chemical compound C[C@H](COC[C@@H](C(O)=O)NC(OCC1c2ccccc2-c2ccccc12)=O)O SXLQXHDUCLWZIE-YJYMSZOUSA-N 0.000 description 1
- SYUXEUKEPIMBRP-XLIONFOSSA-N C[C@H](COC[C@@H](C(OC)=O)NC(OCc1ccccc1)=O)OCc1ccccc1 Chemical compound C[C@H](COC[C@@H](C(OC)=O)NC(OCc1ccccc1)=O)OCc1ccccc1 SYUXEUKEPIMBRP-XLIONFOSSA-N 0.000 description 1
- PXSOUUMARVNCPJ-UVKLAMSESA-N OC([C@H](COCC(C(F)(F)F)OC1OCCCC1)NC(OCC1c2ccccc2-c2c1cccc2)=O)=O Chemical compound OC([C@H](COCC(C(F)(F)F)OC1OCCCC1)NC(OCC1c2ccccc2-c2c1cccc2)=O)=O PXSOUUMARVNCPJ-UVKLAMSESA-N 0.000 description 1
Images


Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K5/00—Peptides containing up to four amino acids in a fully defined sequence; Derivatives thereof
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C229/00—Compounds containing amino and carboxyl groups bound to the same carbon skeleton
- C07C229/02—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton
- C07C229/04—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton being acyclic and saturated
- C07C229/22—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton being acyclic and saturated the carbon skeleton being further substituted by oxygen atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C229/00—Compounds containing amino and carboxyl groups bound to the same carbon skeleton
- C07C229/02—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton
- C07C229/34—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton containing six-membered aromatic rings
- C07C229/36—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton containing six-membered aromatic rings with at least one amino group and one carboxyl group bound to the same carbon atom of the carbon skeleton
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C235/00—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by oxygen atoms
- C07C235/02—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by oxygen atoms having carbon atoms of carboxamide groups bound to acyclic carbon atoms and singly-bound oxygen atoms bound to the same carbon skeleton
- C07C235/04—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by oxygen atoms having carbon atoms of carboxamide groups bound to acyclic carbon atoms and singly-bound oxygen atoms bound to the same carbon skeleton the carbon skeleton being acyclic and saturated
- C07C235/06—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by oxygen atoms having carbon atoms of carboxamide groups bound to acyclic carbon atoms and singly-bound oxygen atoms bound to the same carbon skeleton the carbon skeleton being acyclic and saturated having the nitrogen atoms of the carboxamide groups bound to hydrogen atoms or to acyclic carbon atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C237/00—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups
- C07C237/02—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups having the carbon atoms of the carboxamide groups bound to acyclic carbon atoms of the carbon skeleton
- C07C237/20—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups having the carbon atoms of the carboxamide groups bound to acyclic carbon atoms of the carbon skeleton the carbon skeleton containing six-membered aromatic rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C237/00—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups
- C07C237/28—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups having the carbon atom of at least one of the carboxamide groups bound to a carbon atom of a non-condensed six-membered aromatic ring of the carbon skeleton
- C07C237/30—Carboxylic acid amides, the carbon skeleton of the acid part being further substituted by amino groups having the carbon atom of at least one of the carboxamide groups bound to a carbon atom of a non-condensed six-membered aromatic ring of the carbon skeleton having the nitrogen atom of the carboxamide group bound to hydrogen atoms or to acyclic carbon atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C239/00—Compounds containing nitrogen-to-halogen bonds; Hydroxylamino compounds or ethers or esters thereof
- C07C239/08—Hydroxylamino compounds or their ethers or esters
- C07C239/18—Hydroxylamino compounds or their ethers or esters having nitrogen atoms of hydroxylamino groups further bound to carbon atoms of hydrocarbon radicals substituted by carboxyl groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C259/00—Compounds containing carboxyl groups, an oxygen atom of a carboxyl group being replaced by a nitrogen atom, this nitrogen atom being further bound to an oxygen atom and not being part of nitro or nitroso groups
- C07C259/04—Compounds containing carboxyl groups, an oxygen atom of a carboxyl group being replaced by a nitrogen atom, this nitrogen atom being further bound to an oxygen atom and not being part of nitro or nitroso groups without replacement of the other oxygen atom of the carboxyl group, e.g. hydroxamic acids
- C07C259/10—Compounds containing carboxyl groups, an oxygen atom of a carboxyl group being replaced by a nitrogen atom, this nitrogen atom being further bound to an oxygen atom and not being part of nitro or nitroso groups without replacement of the other oxygen atom of the carboxyl group, e.g. hydroxamic acids having carbon atoms of hydroxamic groups bound to carbon atoms of six-membered aromatic rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C311/00—Amides of sulfonic acids, i.e. compounds having singly-bound oxygen atoms of sulfo groups replaced by nitrogen atoms, not being part of nitro or nitroso groups
- C07C311/01—Sulfonamides having sulfur atoms of sulfonamide groups bound to acyclic carbon atoms
- C07C311/02—Sulfonamides having sulfur atoms of sulfonamide groups bound to acyclic carbon atoms of an acyclic saturated carbon skeleton
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C311/00—Amides of sulfonic acids, i.e. compounds having singly-bound oxygen atoms of sulfo groups replaced by nitrogen atoms, not being part of nitro or nitroso groups
- C07C311/50—Compounds containing any of the groups, X being a hetero atom, Y being any atom
- C07C311/51—Y being a hydrogen or a carbon atom
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D207/00—Heterocyclic compounds containing five-membered rings not condensed with other rings, with one nitrogen atom as the only ring hetero atom
- C07D207/02—Heterocyclic compounds containing five-membered rings not condensed with other rings, with one nitrogen atom as the only ring hetero atom with only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D207/04—Heterocyclic compounds containing five-membered rings not condensed with other rings, with one nitrogen atom as the only ring hetero atom with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members
- C07D207/10—Heterocyclic compounds containing five-membered rings not condensed with other rings, with one nitrogen atom as the only ring hetero atom with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D207/16—Carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/04—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D211/06—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members
- C07D211/08—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms
- C07D211/18—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms
- C07D211/34—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms with hydrocarbon radicals, substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D213/00—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members
- C07D213/02—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members
- C07D213/04—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D213/24—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom with substituted hydrocarbon radicals attached to ring carbon atoms
- C07D213/54—Radicals substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
- C07D213/55—Acids; Esters
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D265/00—Heterocyclic compounds containing six-membered rings having one nitrogen atom and one oxygen atom as the only ring hetero atoms
- C07D265/28—1,4-Oxazines; Hydrogenated 1,4-oxazines
- C07D265/30—1,4-Oxazines; Hydrogenated 1,4-oxazines not condensed with other rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D271/00—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms
- C07D271/02—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms not condensed with other rings
- C07D271/06—1,2,4-Oxadiazoles; Hydrogenated 1,2,4-oxadiazoles
- C07D271/07—1,2,4-Oxadiazoles; Hydrogenated 1,2,4-oxadiazoles with oxygen, sulfur or nitrogen atoms, directly attached to ring carbon atoms, the nitrogen atoms not forming part of a nitro radical
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D271/00—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms
- C07D271/02—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms not condensed with other rings
- C07D271/08—1,2,5-Oxadiazoles; Hydrogenated 1,2,5-oxadiazoles
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D277/00—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings
- C07D277/02—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings
- C07D277/20—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings having two or three double bonds between ring members or between ring members and non-ring members
- C07D277/22—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings having two or three double bonds between ring members or between ring members and non-ring members with only hydrogen atoms, hydrocarbon or substituted hydrocarbon radicals, directly attached to ring carbon atoms
- C07D277/30—Radicals substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D309/00—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only ring hetero atom, not condensed with other rings
- C07D309/02—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only ring hetero atom, not condensed with other rings having no double bonds between ring members or between ring members and non-ring members
- C07D309/08—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only ring hetero atom, not condensed with other rings having no double bonds between ring members or between ring members and non-ring members with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D309/10—Oxygen atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D413/00—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D413/02—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings
- C07D413/12—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings linked by a chain containing hetero atoms as chain links
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D455/00—Heterocyclic compounds containing quinolizine ring systems, e.g. emetine alkaloids, protoberberine; Alkylenedioxy derivatives of dibenzo [a, g] quinolizines, e.g. berberine
- C07D455/03—Heterocyclic compounds containing quinolizine ring systems, e.g. emetine alkaloids, protoberberine; Alkylenedioxy derivatives of dibenzo [a, g] quinolizines, e.g. berberine containing quinolizine ring systems directly condensed with at least one six-membered carbocyclic ring, e.g. protoberberine; Alkylenedioxy derivatives of dibenzo [a, g] quinolizines, e.g. berberine
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D471/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00
- C07D471/12—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00 in which the condensed system contains three hetero rings
- C07D471/14—Ortho-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D498/00—Heterocyclic compounds containing in the condensed system at least one hetero ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D498/12—Heterocyclic compounds containing in the condensed system at least one hetero ring having nitrogen and oxygen atoms as the only ring hetero atoms in which the condensed system contains three hetero rings
- C07D498/14—Ortho-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K11/00—Depsipeptides having up to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K11/02—Depsipeptides having up to 20 amino acids in a fully defined sequence; Derivatives thereof cyclic, e.g. valinomycins ; Derivatives thereof
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K14/00—Peptides having more than 20 amino acids; Gastrins; Somatostatins; Melanotropins; Derivatives thereof
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K5/00—Peptides containing up to four amino acids in a fully defined sequence; Derivatives thereof
- C07K5/04—Peptides containing up to four amino acids in a fully defined sequence; Derivatives thereof containing only normal peptide links
- C07K5/12—Cyclic peptides with only normal peptide bonds in the ring
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K7/50—Cyclic peptides containing at least one abnormal peptide link
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K7/50—Cyclic peptides containing at least one abnormal peptide link
- C07K7/54—Cyclic peptides containing at least one abnormal peptide link with at least one abnormal peptide link in the ring
- C07K7/56—Cyclic peptides containing at least one abnormal peptide link with at least one abnormal peptide link in the ring the cyclisation not occurring through 2,4-diamino-butanoic acid
-
- C—CHEMISTRY; METALLURGY
- C40—COMBINATORIAL TECHNOLOGY
- C40B—COMBINATORIAL CHEMISTRY; LIBRARIES, e.g. CHEMICAL LIBRARIES
- C40B40/00—Libraries per se, e.g. arrays, mixtures
- C40B40/04—Libraries containing only organic compounds
- C40B40/06—Libraries containing nucleotides or polynucleotides, or derivatives thereof
- C40B40/08—Libraries containing RNA or DNA which encodes proteins, e.g. gene libraries
-
- C—CHEMISTRY; METALLURGY
- C40—COMBINATORIAL TECHNOLOGY
- C40B—COMBINATORIAL CHEMISTRY; LIBRARIES, e.g. CHEMICAL LIBRARIES
- C40B40/00—Libraries per se, e.g. arrays, mixtures
- C40B40/04—Libraries containing only organic compounds
- C40B40/10—Libraries containing peptides or polypeptides, or derivatives thereof
-
- Y—GENERAL TAGGING OF NEW TECHNOLOGICAL DEVELOPMENTS; GENERAL TAGGING OF CROSS-SECTIONAL TECHNOLOGIES SPANNING OVER SEVERAL SECTIONS OF THE IPC; TECHNICAL SUBJECTS COVERED BY FORMER USPC CROSS-REFERENCE ART COLLECTIONS [XRACs] AND DIGESTS
- Y02—TECHNOLOGIES OR APPLICATIONS FOR MITIGATION OR ADAPTATION AGAINST CLIMATE CHANGE
- Y02P—CLIMATE CHANGE MITIGATION TECHNOLOGIES IN THE PRODUCTION OR PROCESSING OF GOODS
- Y02P20/00—Technologies relating to chemical industry
- Y02P20/50—Improvements relating to the production of bulk chemicals
- Y02P20/55—Design of synthesis routes, e.g. reducing the use of auxiliary or protecting groups
Definitions
- the present invention relates to an amino acid capable of improving the membrane permeability of a peptide compound, a peptide compound containing the amino acid, and a method for producing them.
- Non-patent Document 1 Non-patent Document 1
- Cyclosporine A and polymyxin B are typical examples of peptide drugs. Focusing on these structures, the characteristic point is that the peptide is a peptide containing some unnatural amino acids represented by N-methyl amino acids, or that it is a cyclic peptide.
- An unnatural amino acid is an amino acid that is not naturally encoded in mRNA.
- these unnatural structural sites or cyclic structures are also included. It is very interesting that the drug contributes to the site of action in the living body and the pharmacokinetics by contributing to the pharmacokinetics.
- Non-Patent Document 1 As a condition for improving membrane permeability (a condition necessary to satisfy drug-likeness) in which a medium-molecular peptide can contribute to improvement in pharmacokinetics, for example, having a cyclic portion, N-substituted amino acid number, Conditions such as the range of the number of amino acid residues and lipophilicity have been reported (Patent Document 1). There is also a report that an N-methyl amino acid which is an unnatural amino acid was focused on and an N-methyl peptide library was produced by translational synthesis (Patent Document 2). Further, in order for the target and the drug to form a complex, the interaction between the target and the drug is important, and the important one among them is the hydrogen bond (Non-Patent Document 3).
- Non-Patent Document 1 A proton donor is essential for forming hydrogen bonds, but it can have a negative effect on membrane permeability.
- N-methylation of amino acids is known to be successful as one of the methods for improving membrane permeability (Patent Document 1).
- Other than N-methylation of amino acids it has been considered that the masking of protons by utilizing an amide bond contained in the main chain affects the membrane permeability (Non-Patent Document 2).
- the improvement of the interaction between the target molecule and the peptide is essential for the creation of a peptide useful for medicine.
- Patent Documents 1 and 2 describe suggestions from the viewpoint of N-methylation of the amide bond site of amino acids, which are the constituent units of peptides, the number of amino acid residues, and lipophilicity, , Consideration on the structural characteristics when a heteroatom represented by an oxygen atom or a nitrogen atom necessary for forming a hydrogen bond which is considered to be advantageous for binding to a target molecule is introduced into the side chain of an amino acid Not done.
- Non-Patent Document 1 describes examples of peptide drugs and cyclization methods for cyclic peptides, but does not suggest a correlation between membrane permeability and amino acid structure.
- Non-Patent Document 2 exemplifies peptide drugs, there is no suggestion that the structural conversion of the amino acids that make up the peptides is linked to their usefulness for drugs, and the consideration of known compounds is only made. However, its versatility in medicine is limited.
- the present invention has been made in view of the situation in which the design of peptide compounds, particularly the effect on the membrane permeability, has been carried out without paying attention to the structural characteristics of each amino acid.
- An object of the present invention is to provide an amino acid that improves the amino acid and a peptide compound containing the amino acid.
- the present inventors have found the following as means for solving the above problems and completed the invention. That is, an amino acid that has a heteroatom necessary for improving the interaction with the target molecule in the side chain of the amino acid and can further mask the proton on the amino acid side chain by intramolecular hydrogen bonding within the same amino acid side chain.
- a peptide compound containing the amino acid was produced, and membrane permeability was evaluated.
- the peptide compound containing the amino acid was compared with a peptide compound containing an amino acid that does not undergo intramolecular hydrogen bonding. Then, it was found that the membrane permeability is excellent.
- the amino acid having a side chain for intramolecular hydrogen bonding is designed so that the proton donor and the proton acceptor can have a pseudo cyclic structure, preferably a 4-membered ring, 5-membered ring, 6-membered ring, or 7-membered ring. It was found that such a structure is effective.
- the number of amino acid residues constituting the peptide optimal for membrane permeation of the peptide compound containing the amino acid the number of N-substituted amino acid residues, the ratio of N-substituted amino acid residues contained in the peptide, the range of cLogP, the aroma, The number of rings was found.
- This invention includes the following in one specific, non-limiting aspect.
- [1] A peptide compound in which two or more amino acids are linked, wherein at least one of the amino acids can form a hydrogen bond in its side chain.
- [2] The peptide compound according to [1], wherein the amino acid capable of forming a hydrogen bond in the side chain can form a pseudo 4- to 7-membered ring in the side chain.
- R 1 is hydrogen, or C 1 -C 6 alkyl, or a group represented by formula 1 or formula 2
- R 2A and R 2B are hydrogen in which R 2A is hydrogen or C 1 -.
- R 2B is hydrogen or C 1 -C 6 alkyl, or R 2A and R 2B are the groups to which they are attached.
- R 2B is hydrogen or C 1 -C 6 alkyl
- R 3 is a single bond or —CHR 4 —
- R 4 is hydrogen, or C 1 -C 4 alkyl, or a group represented by formula 1 or formula 2
- Formula 1 and Formula 2 are respectively the following formulas: Is expressed in the formula, * Indicates the point of attachment
- Q 1 and Q 2 are independently a single bond, C 1 -C 4 alkylene, or C 2 -C 4 heteroalkylene containing one oxygen atom
- a 1 is -O- or -S-
- L 1 is a direct group which may be substituted with one or more substituents selected from the group consisting of fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- a chain C 1 -C 3 alkylene, A 2 is a single bond, —O—, or —S—, L 2 is a single bond or is one or more substituents selected from the group consisting of fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- a linear C 1 -C 3 alkylene which may be substituted X is —OH, —NR Z1 R Z2 , or —CONR Z1 R Z2 , or contains 1 to 3 heteroatoms, optionally substituted by oxo or one or more halogen 5 ⁇ 6-membered saturated or unsaturated heterocyclyl;
- R Z1 and R Z2 are independently selected from the group consisting of hydrogen, —OH, C 1 -C 4 alkyl, and C 1 -C 4 alkylsulfonyl
- Y is —OH, C 1 -C 4 alkylsulfonylamino, —NR Z3 R Z4 , or —CONR Z3 R Z4 , or contains 1 to 3 heteroatoms and is substituted by oxo or halogen.
- R Z3 and R Z4 are independently selected from the group consisting of hydrogen, —OH, C 1 -C 4 alkyl, and C 1 -C 4 alkylsulfonyl, Z is hydrogen or C 1 -C 4 alkyl, However, when A 2 is —O— or —S—, Z is not hydrogen,
- the amino acid represented by formula A contains at least one group represented by formula 1 or formula 2.
- R 1 is a group represented by formula 1 or formula 2
- R 2A and R 2B are independently hydrogen, or C 1 -C 6 alkyl, or R 2A and R 2B are 4-6 membered together with the carbon atom to which they are attached.
- R 3 is a single bond or —CHR 4 —
- R 1 forms a 4- to 6-membered heterocycle together with the nitrogen atom to which R 1 is bonded, R 2A , and the carbon atom to which R 2A is bonded, and the heterocycle has the formula 1 or Having a group represented by formula 2,
- R 2B is hydrogen or C 1 -C 6 alkyl
- R 3 is a single bond or —CHR 4 —,
- R 1 is hydrogen or C 1 -C 6 alkyl
- R 2A is a group represented by formula 1 or formula 2
- R 2B is hydrogen or C 1 -C 6 alkyl
- R 3 is a single bond or —CHR 4 —
- R 1 is hydrogen or C 1 -C 6 alkyl
- R 2A and R 2B are independently hydrogen, or C 1 -C 6 alkyl
- R 2A and R 2B are 4-6 membered together with the carbon atom to which they are attached.
- R 3 is —CHR 4 —
- R 4 is a group represented by formula 1 or formula 2,
- Q 1 is C 1 -C 4 alkylene
- a 1 is —O— or —S—
- L 1 is a straight chain which may be substituted with one or more substituents selected from the group consisting of C 1 -C 2 alkyl groups, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- X is -OH
- Q 1 is C 1 -C 4 alkylene
- a 1 is —O— or —S—
- L 1 is a straight chain which may be substituted with one or more substituents selected from the group consisting of C 1 -C 2 alkyl groups, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- X is -CONR Z1 R Z2
- R Z1 is C 1 -C 4 alkyl
- R Z2 is hydrogen
- the peptide compound according to any one of [3] to [7].
- Q 1 is C 1 -C 4 alkylene
- a 1 is —O— or —S—
- L 1 is a straight chain which may be substituted with one or more substituents selected from the group consisting of C 1 -C 2 alkyl groups, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- X is a 5-6 membered saturated or unsaturated heterocyclyl containing 1-3 heteroatoms and optionally substituted by oxo or by one or more halogens
- the peptide compound according to any one of [3] to [7].
- Q 1 is a single bond
- a 1 is —O— or —S—
- L 1 is a straight chain which may be substituted with one or more substituents selected from the group consisting of C 1 -C 2 alkyl groups, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O).
- X is -OH
- [12] The peptide compound according to any one of [3] to [11], wherein Q 1 is CH 2 —.
- [13] The peptide compound according to any one of [3] to [12], wherein A 1 is —O—.
- L 1 is —CH 2 —, —(CH 2 ) 2 —, —(CH 2 ) 3 —, —CH 2 CH(CH 3 )—, —CH 2 CH(CF 3 )—, —CH 2 C(CH 3 ) 2 —, —(CH 2 ) 2 CH(CH 3 )—, —CH 2 C(CH 3 ) 2 CH 2 —, —(CH 2 ) 2 C(CH 3 ) 2 —, and
- Q 2 is C 1 -C 4 alkylene, L 2 is a single bond, Y is -CONR Z3 R Z4 , A 2 is -O-, or -S-, Z is C 1 -C 4 alkyl, The peptide compound according to any one of [3] to [7].
- Q 2 is C 1 -C 4 alkylene, L 2 is a single bond, Y is -CONR Z3 R Z4 , A 2 is a single bond, Z is hydrogen, The peptide compound according to any one of [3] to [7].
- Q 2 is C 1 -C 4 alkylene, L 2 is a single bond, Y is C 1 -C 4 alkylsulfonylaminocarbonyl, A 2 is -O-, or -S-, Z is C 1 -C 4 alkyl, The peptide compound according to any one of [3] to [7].
- Q 2 is C 1 -C 4 alkylene, L 2 is a single bond, Y is -OH, A 2 is -O-, or -S-, Z is C 1 -C 4 alkyl, The peptide compound according to any one of [3] to [7].
- [29] The peptide compound according to any one of [1] to [28], wherein the ratio of the number of N-substituted amino acids to the total number of amino acids is 30% or more.
- [34] A group consisting of: The amino acid according to [33], which is selected from: [35] A protected amino acid, wherein the amino group and/or the carboxyl group contained in the amino acid according to [33] or [34] is protected by a protecting group.
- the amino-protecting group is selected from the group consisting of Fmoc group, Boc group, Cbz group, Alloc group, nosyl group, dinitronosyl group, t-Bu group, trityl group, and cumyl group, and/or The protected amino acid according to [35], wherein the protective group for the carboxyl group is selected from the group consisting of a methyl group, an allyl group, a t-Bu group, a trityl group, a cumyl group, a methoxytrityl group, and a benzyl group.
- FIG. 6 is a diagram showing a relationship between ClogP/totalAA and Papp when a cyclic peptide compound having no Trp side chain is used as a population.
- FIG. 3 is a diagram showing a relationship between ClogP/totalAA and Papp when a cyclic peptide compound having a Trp side chain is used as a population. It is a figure showing the relation between ARC of a cyclic peptide compound and cell membrane permeability. For example, it is a diagram of a potential energy surface (Potential energy surface) in the side chain partial structure of Ser(NMe-Aca).
- FIG. 1 it is a diagram showing the conformational distribution of crystal structures (Crystal structures) using the results of X-ray structural analysis registered in CSD in the side chain partial structure of Ser(NMe-Aca).
- FIG. 1 it is a diagram of a potential energy curved surface in a side chain partial structure of Ser(EtOH), bAla(3R-MeOEtOH), and bAla(2S-MeOEtOH).
- the conformational distribution of the crystal structure was calculated. It is the figure shown.
- alkyl is a monovalent group derived from an aliphatic hydrocarbon by removing one arbitrary hydrogen atom, and refers to a hetero atom (an atom other than carbon and hydrogen atoms) in the skeleton. ) Or unsaturated carbon-carbon bonds and has a subset of hydrocarbyl or hydrocarbon radical structures containing hydrogen and carbon atoms.
- the alkyl group includes a straight chain or a branched chain.
- the alkyl group is an alkyl group having 1 to 20 carbon atoms (C 1 -C 20 , hereinafter "C p -C q "means that the carbon number is p to q).
- a C 1 -C 6 alkyl group a C 1 -C 5 alkyl group, a C 1 -C 4 alkyl group, a C 1 -C 3 alkyl group and the like can be mentioned.
- the alkyl include methyl, ethyl, propyl, butyl, pentyl, hexyl, isopropyl, tert-butyl group, sec-butyl group, 1-methylpropyl group, 1,1-dimethylpropyl group and 2,2.
- heteroalkyl means a group containing preferably 1 to 5 heteroatoms in the skeleton of the “alkyl”, preferably C 2 -C 10 heteroalkyl, C 2 -C 6 hetero. Examples thereof include alkyl. Specific examples of the heteroalkyl include —CH 2 OCH 3 , —CH 2 OCH 2 CH 3 , —CH(CH 3 )OCH 3 , and —CH 2 CH 2 N(CH 3 ) 2 .
- halogen atom in the present specification includes F, Cl, Br, and I, and F or Cl is preferably exemplified.
- fluoroalkyl in the present specification means a group in which one or more hydrogen atoms of the above “alkyl” are substituted with a fluorine atom, preferably C 1 -C 6 fluoroalkyl, C 1 -C 4 fluoro Alkyl, C 1 -C 2 fluoroalkyl and the like can be mentioned.
- fluoroalkyl include trifluoromethyl group, 2,2,2-trifluoroethyl group, pentafluoroethyl group, 2,2,3,3-tetrafluoropropyl group, heptafluoropropyl group, trifluoromethoxy group.
- 2,2,2-trifluoroethoxy group pentafluoroethoxy group, 2,2,3,3-tetrafluoropropoxy group, heptafluoropropoxy group and the like.
- alkylsulfonyl used herein means a sulfonyl group to which the above “alkyl” is bonded (ie, —SO 2 -alkyl).
- alkylsulfonyl include C 1 -C 6 alkylsulfonyl, C 1 -C 4 alkylsulfonyl and the like, and specific examples include methylsulfonyl, ethylsulfonyl, n-propylsulfonyl, i-propylsulfonyl and the like. Be done.
- alkylsulfonylamino means a group in which one hydrogen atom of an amino group (—NH 2 ) is substituted with the above “alkylsulfonyl”.
- alkylsulfonylamino include C 1 -C 6 alkylsulfonylamino, C 1 -C 4 alkylsulfonylamino and the like, and specific examples include methylsulfonylamino, ethylsulfonylamino, n-propylsulfonylamino, i. -Propylsulfonylamino and the like.
- alkenyl group in the present specification is a monovalent group having at least one double bond (two adjacent sp 2 carbon atoms). Depending on the arrangement of double bonds and substituents (if present), the double bond geometry can assume the Mais (E) or sixteen (Z), cis or trans configurations.
- alkenyl group include linear or branched ones, and include linear chains containing an internal olefin and the like. Preferred is a C 2 -C 10 alkenyl group, and more preferred is a C 2 -C 6 alkenyl group.
- alkenyl examples include, for example, vinyl group, allyl group, 1-propenyl group, 2-propenyl group, 1-butenyl group, 2-butenyl group (including cis and trans), and 3-butenyl group. , A pentenyl group, a hexenyl group and the like.
- alkynyl is a monovalent group having at least one triple bond (two adjacent SP carbon atoms). Included are straight or branched chain alkynyl groups, including internal alkylene. A C 2 -C 10 alkynyl group is preferable, and a C 2 -C 6 alkynyl group is more preferable. Specific examples of alkynyl include an ethynyl group, a 1-propynyl group, a propargyl group, a 3-butynyl group, a pentynyl group and a hexynyl group.
- cycloalkyl means a saturated or partially saturated cyclic monovalent aliphatic hydrocarbon group and includes a monocycle, a bicyclo ring and a spiro ring. Preferable one is C 3 -C 10 cycloalkyl.
- the cycloalkyl group may be partially unsaturated. Specific examples of cycloalkyl include cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl, cycloheptyl, cyclooctyl, bicyclo[2.2.1]heptyl and the like.
- aryl means a monovalent aromatic hydrocarbon ring, and preferably C 6 -C 10 aryl is mentioned. Specific examples of the aryl include phenyl and naphthyl.
- heteroaryl refers to one aromatic ring containing preferably 1 to 5 heteroatoms in the atoms constituting the ring (also referred to as “in the ring” in the present specification).
- a valent group is meant and may be partially saturated.
- the ring may be monocyclic or two fused rings (eg, bicyclic heteroaryl fused with benzene or monocyclic heteroaryl).
- the number of atoms constituting the ring is preferably 5-10 (5-10 membered heteroaryl).
- heteroaryl examples include imidazolyl, thiazolyl, oxadiazolyl, thiadiazolyl, oxadiazolonyl, thiadiazolonyl, tetrazolyl, pyridyl, indolyl and the like.
- “having a hetero atom in the ring” means containing a hetero atom in the atoms constituting the ring, and examples of such a ring include an aromatic heterocycle such as pyridine and piperidine. , Non-aromatic heterocycles such as morpholine, pyrrolidine, or azetidine. Further, when the hetero atom is an oxygen atom, it is described as “having an oxygen atom in the ring”.
- the “heterocyclic group” means a group having at least one hetero atom (eg, N, O, S, etc.) in the ring, and is also called a saturated heterocyclyl group or an unsaturated heterocyclyl group.
- the hetero atom is preferably N or O, the number of hetero atoms is preferably 1 or 2, and is preferably a 4- to 6-membered ring.
- the heterocyclic group may be substituted with an alkyl group, a fluoroalkyl group, oxo or a halogen atom.
- heterocyclic group examples include a pyridyl group, a piperidino group, a morpholino group, a pyrrolidino group, an oxadiazolonyl group, and an azetidinyl group.
- arylalkyl is a group containing both aryl and alkyl, and means, for example, a group in which at least one hydrogen atom of the alkyl is substituted with aryl, and preferably "C" 5 -C 10 aryl C 1 -C 6 alkyl".
- benzyl and the like can be mentioned.
- alkylene means a divalent group derived by removing any one hydrogen atom from the above “alkyl”.
- alkylene include C 1 -C 2 alkylene, C 1 -C 3 alkylene, C 1 -C 4 alkylene, C 1 -C 5 alkylene and C 1 -C 6 alkylene.
- alkylene examples include —CH 2 —, —(CH 2 ) 2 —, —(CH 2 ) 3 —, CH(CH 3 )CH 2 —, —C(CH 3 ) 2 —, —(CH 2 ) 4 —, CH(CH 3 )CH 2 CH 2 —, —C(CH 3 ) 2 CH 2 —, —CH 2 CH(CH 3 )CH 2 —, —CH 2 C(CH 3 ) 2 —, — CH 2 CH 2 CH(CH 3 )—, —(CH 2 ) 5 —, —(CH 2 ) 6 — and the like can be mentioned.
- heteroalkylene means a divalent group derived by removing one arbitrary hydrogen atom from the above “heteroalkyl”, and is a C 2 -C 6 heteroalkylene, C 2 -C 5 Heteroalkylene, C 2 -C 4 heteroalkylene, C 2 -C 3 heteroalkylene, C 2 heteroalkylene and the like can be mentioned.
- specific examples of the heteroalkylene containing an oxygen atom as a hetero atom in the group include —CH 2 O—, —OCH 2 —, —CH 2 OCH 2 —, —OCH 2 CH 2 —, —CH 2 CH 2 O- and the like.
- the “arylene” means a divalent group derived by removing one arbitrary hydrogen atom from the aryl.
- Arylene may be a single ring or a condensed ring.
- the number of atoms constituting the ring is not particularly limited, but is preferably 6 to 10 (C 6 -C 10 arylene).
- Specific examples of the arylene include phenylene and the like.
- heteroarylene means a divalent group derived by removing one arbitrary hydrogen atom from the above heteroaryl.
- the heteroarylene may be a single ring or a condensed ring.
- the number of atoms constituting the ring is not particularly limited, but is preferably 5 to 10 (5- to 10-membered heteroarylene).
- Specific examples of the heteroarylene include imidazoldiyl, pyridinediyl, oxadiazolediyl, thiazolediyl, thiadiazolediyl and the like. ..
- the “condensed ring structure” refers to a cyclic structure in which a plurality of rings share two or more atoms in a cyclic compound having two or more rings.
- the “condensed ring structure with two or more aromatic rings” refers to a cyclic compound having two or more aromatic rings, in which a plurality of aromatic rings share two or more atoms.
- examples of the condensed ring structure include an indole skeleton, a benzofuran skeleton, a benzimidazole skeleton, a quinoline skeleton, and a bicyclo[4.4.0]decane.
- the substituent is, for example, an alkyl group, a fluoroalkyl group, an alkoxy group, a fluoroalkoxy group, an alkenyl group, or an alkenyloxy group.
- heterocycle in the present specification means a non-aromatic monovalent or divalent heterocycle having preferably 1 to 5 heteroatoms in the atoms constituting the ring.
- the heterocycle may have a double and/or triple bond in the ring, a carbon atom in the ring may be oxidized to form a carbonyl, and may be a monocycle, a condensed ring or a spiro ring.
- the number of atoms constituting the ring is preferably 3 to 12 (3 to 12 membered heterocycle), more preferably 4 to 7 (4 to 7 membered heterocycle), and further preferably 5 to 6 ( 5-6 membered heterocycle).
- heterocycle examples include piperazine, pyrrolidine, piperidine, morpholine, homomorpholine, (R)-hexahydropyrrolo[1,2-a]pyrazine, (S)-hexahydropyrrolo[1,2-a ] Pyrazine, 3-oxopiperazine, 2-oxopyrrolidine, azetidine, 2-oxoimidazolidine, oxetane, dihydrofuran, tetrahydrofuran, dihydropyran, tetrahydropyran, tetrahydropyridine, thiomorpholine, pyrazolidine, imidazoline, oxazolidine, isoxazolidine, thiazolidine , Imidazolidine, isothiazolidine, thiadiazolidine, oxazolidone, benzodioxane, benzoxazoline, dioxolane, dioxane and tetrahydrothiopyran
- the present invention provides a peptide compound in which two or more amino acids are linked, wherein at least one amino acid (specifically, for example, one, two, three, four, or more) Is a amino acid capable of forming a hydrogen bond in its side chain.
- the “peptide compound” in the present invention includes a linear or cyclic peptide compound in which two or more amino acids are linked.
- the cyclic peptide compound has the same meaning as "a peptide compound having a cyclic portion".
- amino acid in the present specification includes natural amino acids and unnatural amino acids.
- the “natural amino acid” in the present specification means Gly, Ala, Ser, Thr, Val, Leu, Ile, Phe, Tyr, Trp, His, Glu, Asp, Gln, Asn, Cys, Met, Lys, Arg, Pro.
- the unnatural amino acid is not particularly limited, but examples thereof include ⁇ -amino acids, D-type amino acids, N-substituted amino acids, ⁇ , ⁇ -disubstituted amino acids, amino acids whose side chains differ from natural amino acids, and hydroxycarboxylic acids. As the amino acid in the present specification, any configuration is allowed.
- the selection of the side chain of the amino acid is not particularly limited, but in addition to a hydrogen atom, for example, an alkyl group, an alkenyl group, an alkynyl group, an aryl group, a heteroaryl group, an aralkyl group, a cycloalkyl group, a spiro-bonded cycloalkyl group.
- a hydrogen atom for example, an alkyl group, an alkenyl group, an alkynyl group, an aryl group, a heteroaryl group, an aralkyl group, a cycloalkyl group, a spiro-bonded cycloalkyl group.
- Freely selected from the base may have a substituent, and the substituent is not limited, and any substituent including a halogen atom, an O atom, an S atom, an N atom, a B atom, a Si atom, or a P atom can be given.
- One or two or more
- examples thereof include an optionally substituted alkyl group, alkenyl group, alkynyl group, aryl group, heteroaryl group, aralkyl group, cycloalkyl group and the like.
- the amino acid in the present specification may have a side chain “which can form an intramolecular hydrogen bond” described later.
- the amino acid in the present specification may be a compound having a carboxy group and an amino group in the same molecule (even in this case, imino acids such as proline and hydroxyproline are also amino acids). included).
- substituent derived from halogen examples include fluoro (-F), chloro (-Cl), bromo (-Br), iodo (-I) and the like.
- oxy examples include alkoxy, cycloalkoxy, alkenyloxy, alkynyloxy, aryloxy, heteroaryloxy, aralkyloxy and the like.
- carbonyl examples include formyl (—C ⁇ O—H), alkylcarbonyl, cycloalkylcarbonyl, alkenylcarbonyl, alkynylcarbonyl, arylcarbonyl, heteroarylcarbonyl, aralkylcarbonyl and the like. ..
- Examples of carbonyloxy include alkylcarbonyloxy, cycloalkylcarbonyloxy, alkenylcarbonyloxy, alkynylcarbonyloxy, arylcarbonyloxy, heteroarylcarbonyloxy, aralkylcarbonyloxy and the like. ..
- thiocarbonyl examples include alkylthiocarbonyl, cycloalkylthiocarbonyl, alkenylthiocarbonyl, alkynylthiocarbonyl, arylthiocarbonyl, heteroarylthiocarbonyl, aralkylthiocarbonyl and the like.
- carbonylthio examples include alkylcarbonylthio, cycloalkylcarbonylthio, alkenylcarbonylthio, alkynylcarbonylthio, arylcarbonylthio, heteroarylcarbonylthio, aralkylcarbonylthio and the like. ..
- aminocarbonyl examples include alkylaminocarbonyl, cycloalkylaminocarbonyl, alkenylaminocarbonyl, alkynylaminocarbonyl, arylaminocarbonyl, heteroarylaminocarbonyl, aralkylaminocarbonyl and the like.
- Examples of carbonylamino include alkylcarbonylamino, cycloalkylcarbonylamino, alkenylcarbonylamino, alkynylcarbonylamino, arylcarbonylamino, heteroarylcarbonylamino, aralkylcarbonylamino and the like. ..
- compounds in which the H atom bonded to the N atom in —NH—C ⁇ O—R is further substituted with alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, or aralkyl are included.
- Examples of oxycarbonylamino include alkoxycarbonylamino, cycloalkoxycarbonylamino, alkenyloxycarbonylamino, alkynyloxycarbonylamino, aryloxycarbonylamino, heteroaryloxycarbonylamino, aralkyloxy. Carbonylamino and the like can be mentioned. In addition to these, compounds in which the H atom bonded to the N atom in —NH—C ⁇ O—OR is further substituted with alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, or aralkyl are included.
- Examples of sulfonylamino include alkylsulfonylamino, cycloalkylsulfonylamino, alkenylsulfonylamino, alkynylsulfonylamino, arylsulfonylamino, heteroarylsulfonylamino, aralkylsulfonylamino and the like.
- compounds in which the H atom bonded to the N atom in —NH—SO 2 —R is further substituted with alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, or aralkyl are included.
- aminosulfonyl examples include alkylaminosulfonyl, cycloalkylaminosulfonyl, alkenylaminosulfonyl, alkynylaminosulfonyl, arylaminosulfonyl, heteroarylaminosulfonyl, aralkylaminosulfonyl and the like.
- compounds in which the H atom bonded to the N atom in —SO 2 —NHR is further substituted with alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, or aralkyl are included.
- sulfamoylamino examples include alkylsulfamoylamino, cycloalkylsulfamoylamino, alkenylsulfamoylamino, alkynylsulfamoylamino, arylsulfamoylamino, hetero Arylsulfamoylamino, aralkylsulfamoylamino and the like can be mentioned.
- the two H atoms bonded to the N atom in —NH—SO 2 —NHR are substituents independently selected from the group consisting of alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, and aralkyl. They may be substituted, and these two substituents may form a ring.
- thio are selected from alkylthio, cycloalkylthio, alkenylthio, alkynylthio, arylthio, heteroarylthio, aralkylthio and the like.
- sulfinyl examples include alkylsulfinyl, cycloalkylsulfinyl, alkenylsulfinyl, alkynylsulfinyl, arylsulfinyl, heteroarylsulfinyl, aralkylsulfinyl and the like.
- sulfonyl examples include alkylsulfonyl, cycloalkylsulfonyl, alkenylsulfonyl, alkynylsulfonyl, arylsulfonyl, heteroarylsulfonyl, aralkylsulfonyl and the like.
- secondary amino examples include alkylamino, cycloalkylamino, alkenylamino, alkynylamino, arylamino, heteroarylamino, aralkylamino and the like.
- tertiary amino are each independently selected from alkyl(aralkyl)amino and the like, alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, aralkyl and the like. , And an amino group having any two substituents, and these two optional substituents may form a ring.
- Examples of substituted amidino include three substituents R, R', and R" on the N atom are alkyl, cycloalkyl, alkenyl, alkynyl, aryl, hetero. Examples include groups independently selected from aryl and aralkyl, such as alkyl(aralkyl)(aryl)amidino.
- R, R', R", and R"' are alkyl, cycloalkyl, alkenyl, alkynyl, aryl. , A heteroaryl, an aralkyl, a group independently selected from the above, or a group in which these form a ring.
- aminocarbonylamino examples include R, R′, and R′′ selected from hydrogen atom, alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, and aralkyl. Examples include groups independently selected from each other, or groups forming a ring.
- substituent derived from the B atom examples include boryl (-BR(R')) and dioxyboryl (-B(OR)(OR')). These two substituents R and R'are each independently selected from alkyl, cycloalkyl, alkenyl, alkynyl, aryl, heteroaryl, aralkyl, etc., or they may form a ring. ..
- an amino acid means a molecule having at least one amino group and at least one carboxyl group in the molecule, but in the present specification.
- a hydroxycarboxylic acid having a hydroxyl group and a carboxyl group in the molecule can also be included in the amino acid of the present invention.
- hydroxycarboxylic acid may be referred to as hydroxyamino acid.
- ⁇ amino acid means that the amino group and the carboxyl group in the amino acid molecule have the same carbon bonded to each other, and the substituent on the carbon is called the side chain of the amino acid.
- the amino group, the carboxyl group, and the series of carbon atoms to which they are bound in the amino acid molecule are called the main chain of the amino acid.
- the main chain amino group of an amino acid may be unsubstituted (NH 2 group) or may be substituted (that is, —NHR group: R is an optionally substituted alkyl, alkenyl, alkynyl, aryl, Represents a heteroaryl, aralkyl or cycloalkyl, and a carbon chain bonded to the N atom and a carbon atom at the ⁇ -position may form a ring like proline).
- An amino acid in which the main chain amino group is substituted is referred to as “N-substituted amino acid” in the present specification.
- N-substituted amino acid in the present specification is preferably N-alkyl amino acid, N—C 1 -C 6 alkyl amino acid, N—C 1 -C 4 alkyl amino acid, N-methyl amino acid, “intramolecular hydrogen bond Examples include, but are not limited to, N-substituted amino acids having "formable" side chains.
- amino acid constituting the peptide compound in the present specification includes all isotopes corresponding to each.
- An “amino acid” isotope is one in which at least one atom has been replaced with an atom having the same atomic number (proton number) but different mass numbers (sum of protons and neutrons).
- isotopes contained in the "amino acid" constituting the peptide compound of the present invention include a hydrogen atom, a carbon atom, a nitrogen atom, an oxygen atom, a phosphorus atom, a sulfur atom, a fluorine atom, a chlorine atom, and the like, respectively, 2 H, 3 H, 13 C, 14 C, 15 N, 17 O, 18 O, 32 P, 35 S, 18 F, 36 Cl and the like are included.
- the “linear peptide compound” in the present invention is a compound formed by linking a natural amino acid and/or an unnatural amino acid with an amide bond or an ester bond, and as long as it is a compound having no cyclic portion, It is not particularly limited.
- the total number of natural amino acids or non-natural amino acids constituting the linear peptide compound is 1, 2, 3, 4, 5, 6, 7, 8, 9, 10, 11, 12, 13, 14, 15, 20. , 25, 30 and a preferable range is 6 to 20, 7 to 19, 7 to 18, 7 to 17, 7 to 16, 7 to 15, 8 to 14, 9 It is up to 13.
- the “cyclic peptide compound” in the present invention is formed by connecting a natural amino acid and/or an unnatural amino acid with an amide bond or an ester bond, and is not particularly limited as long as it is a compound having a cyclic portion.
- the total number of natural amino acids or unnatural amino acids constituting the cyclic peptide compound is 1, 2, 3, 4, 5, 6, 7, 8, 9, 10, 11, 12, 13, 14, 15, 20, 25. , 30 can be used, and the preferred range is 6 to 20, 7 to 19, 7 to 18, 7 to 17, 7 to 16, 7 to 15, 8 to 14, 9 to 13. It is an individual.
- the “cyclic portion” of a peptide compound means a cyclic portion formed by connecting two or more amino acid residues.
- the “linear portion” used when referring to the partial structure of the cyclic peptide compound is a portion that is not included in the main chain structure of the cyclic portion and has at least one on the chain of the portion. It has one amide bond and/or ester bond.
- the number of amino acids constituting the cyclic portion of the cyclic peptide compound in the present specification is not limited, for example, 2 or more, 3 or more, 4 or more, 6 or more, 7 or more, 8 or more, 9 or more, 10 or more, 11 or more, 12 or more, 30 or less, 20 or less, 18 or less, 16 or less, 15 or less, 14 or less, 13 or less, 12 or less, 11 or less, 7, 8, 9, 10, 11, 12, 13, 13, 14, 15, and 16 Is mentioned.
- the number of amino acids forming the cyclic portion is preferably 2 to 30, 2 to 15, or 5 to 15 and 5 to 14, 7 to 14, or 8 to 14 is more preferable, 8 to 13, 9 to 13, 8 to 12, 8 to 11, or 9 to 12 is further preferable, and 9 to 11 is particularly preferable.
- the number of amino acids in the linear portion is preferably 0 to 8, more preferably 0 to 5, and even more preferably 0 to 3.
- the “linear portion” in the present specification may include natural amino acids and unnatural amino acids (including amino acids that have been chemically modified or skeleton-converted).
- the peptide compound or cyclic peptide compound in the present specification is preferably one that does not have an ester bond in the cyclic portion or the linear portion.
- the cyclic peptide compound in the present specification does not have at least one selected from the group consisting of (A) to (C) below in the side chain of the peptide moiety of the cyclic portion or linear portion. It may be a cyclic peptide compound: (A) Indole skeleton; (B) a condensed ring structure with two or more aromatic rings; (C) an unsubstituted hydroxyphenyl group. Among these, a cyclic peptide compound having no (A) or (C) in the side chain of the cyclic part is preferable, and a cyclic peptide compound having no (B) or (C) in the side chain of the cyclic part is more preferable.
- the “condensed ring structure of two or more aromatic rings” may be a “condensed ring structure”.
- the cyclic peptide compounds herein can have a molecular weight of 500-2000.
- the cyclic peptide compound in the present specification is at least one selected from the group consisting of (A) methylthio group and (B) thiol group in the side chain of the peptide moiety of the cyclic moiety or the linear moiety. Or a cyclic peptide compound having neither or both.
- the OH group of the carboxyl group of the main chain in the first amino acid is the nitrogen of the amino group of the main chain in the second amino acid. Substitution with atomic moieties forms peptide compounds.
- the OH group of the carboxyl group of the main chain in the first amino acid is the oxygen of the hydroxyl group of the main chain in the second hydroxyamino acid. Substitution with atomic moieties forms peptide compounds.
- a peptide compound when a peptide compound is formed by linking a natural amino acid and/or an unnatural amino acid with an amide bond or an ester bond, two or more amino acids are consecutively connected by an amide bond and/or an ester bond. Are bound to form a peptide compound.
- a site linked by an amide bond and/or an ester bond of a peptide compound may be referred to as a “peptide site” in the present specification.
- the bonding mode of the cyclization part is not limited to an amide bond or an ester bond.
- the bonding mode of the cyclized portion include an amide bond, a carbon-carbon bond, a disulfide bond, an ester bond, a thioester bond, a thioether bond, a lactam bond, a bond via a triazole structure, and a bond via a fluorophore structure.
- the cyclic peptide compound in the present specification preferably has an amide bond in the cyclized portion.
- the “bonding mode of the cyclized portion” refers to the bonding mode of the site where cyclization is formed by the cyclization reaction.
- the “peptide compound” in the present invention in addition to the above-mentioned condition of the total number of natural amino acids and unnatural amino acids, or alone, at least two N-substituted amino acids (preferably 2, 3, 4, 5, 6, 7,8,9,10,11,12,13,14,15,20,25,30, particularly preferably 5,6 or 7, with a preferred range of 2 to 30, 3 to 30, 6 to (20, 7 to 19, 7 to 18, 7 to 17, 7 to 16, 7 to 15, 8 to 14, 9 to 13), and at least one amino acid that is not N-substituted It can be a linear or cyclic peptide, including.
- N-substituted examples include, but are not limited to, substitution of a hydrogen atom bonded to an N atom with a methyl group, an ethyl group, a propyl group, a butyl group, a hexyl group, or the like.
- the N-substituted amino acid include amino acids in which an amino group contained in a natural amino acid is N-methylated, N-ethylated, N-propylated, N-butylated, or N-pentylated. -Methyl amino acid, N-ethyl amino acid, N-propyl amino acid, N-butyl amino acid, N-pentyl amino acid.
- N-substituted Converting an N-unsubstituted amino acid to an N-substituted amino acid is called N-substituted, and is sometimes called N-alkylation, N-methylation, or N-ethylation.
- the ratio of the number of N-substituted amino acids contained in the peptide compound in the present invention is 30% or more, 40% or more, 50% or more, 60% or more, 70% or more, 80% or more of the total number of amino acids constituting the peptide compound. It is illustrated.
- the number of N-substituted amino acids contained in the peptide moiety of the cyclic peptide compound in the present specification is preferably 2 or more or 3 or more, more preferably 4 or more, 5 or more, or 6 or more, and 7 or more. Is more preferable, 8 or more is particularly preferable, and 20 or less, 15 or less, 14 or less, 13 or less, 12 or less, 10 or less, 9 or less are preferably exemplified.
- the number of N-substituted amino acids contained in the cyclic peptide compound in the present specification is 30% or more, 40% or more, 50% or more, 60% or more, 70% or more, 80% or more of the number of amino acids constituting the cyclic portion.
- the N-substituted amino acid referred to herein may be preferably an N-alkyl amino acid, more preferably an N-methyl amino acid. That is, in one non-limiting aspect, the number of N-substituted amino acids is exemplified as the number of N-alkyl amino acids or the number of N-methyl amino acids. When the number of amino acids constituting the cyclic portion is 8, the number of N-substituted amino acids is preferably 3-7. When the number of amino acids constituting the cyclic portion is 9, the number of N-substituted amino acids is preferably 3-8.
- the number of amino acids constituting the cyclic portion is 10
- the number of N-substituted amino acids is preferably 3-8.
- the number of N-substituted amino acids is preferably 4-9.
- the number of amino acids constituting the cyclic portion is 12, the number of N-substituted amino acids is preferably 4-10.
- the number of amino acids constituting the cyclic portion is 13, the number of N-substituted amino acids is preferably 4-11.
- the number of amino acids constituting the cyclic portion is 14, the number of N-substituted amino acids is preferably 5-12.
- amino acid that constitutes a peptide compound may be referred to as “amino acid residue”.
- the “peptide compound” in the present specification may include a pharmaceutically acceptable salt thereof or a solvate thereof.
- side chain is used in a context such as a side chain of an amino acid or a side chain of a cyclic portion of a cyclic peptide compound, and means a portion not included in each main chain structure.
- the “number of amino acids” refers to the number of amino acid residues (amino acid units) constituting a peptide compound, and includes the amide bond, the ester bond, and the bond of the cyclization part that connect amino acids. It means the number of amino acid units generated upon cleavage.
- the pKa used in the present specification refers to the actually measured pKa.
- a pKa value obtained by ADMET Predictor described later is referred to as a calculated pKa.
- the Basic pKa used in the present specification refers to the actually measured Basic pKa unless otherwise indicated.
- the Basic pKa value obtained by ADMET Predictor described later is referred to as Basic calculated pKa.
- ⁇ pKa and Basic pKa can be obtained by the usual method. For example, it can be measured by the method described in Experimental Chemistry Course 5 "Thermal Measurement and Equilibrium", page 460 (edited by The Chemical Society of Japan, published by Maruzen Co., Ltd.). More specifically, it can be measured by the method described in Reference Example 3-1. Also, when the pKa value of the side chain of the amino acid to be measured and the Basic pKa value are difficult to obtain due to the influence of other functional groups, only the pKa of the target functional group and Basic pKa can be measured, It is possible to measure by appropriately protecting other functional groups with a protecting group or the like.
- acidic side chain means a side chain having a pKa of 10 or less
- basic side chain means a side chain having a Basic pKa of 4 or more.
- a side chain having a pKa of more than 10 and a side chain having a Basic pKa of less than 4 are defined as neutral side chains.
- the peptide compound or cyclic peptide compound in the present specification may contain an amino acid having an acidic side chain that does not hydrogen bond intramolecularly in addition to an amino acid having a side chain capable of intramolecular hydrogen bonding. it can.
- the pKa of the acidic side chain may be 3.5-10.
- the pKa of the acidic side chain is preferably 3.5 or more, more preferably 3.9 or more, more preferably 4.5 or more, more preferably 5.0 or more, more preferably 5.5 or more, More preferably, it is 5.7 or more.
- the Basic pKa is preferably 10 or less.
- the preferable pKa range is preferably 3.5 to 10, 3.9 to 10, 4.5 to 10, 5.0 to 10, or 5.5 to 10, 5.7 to 10.
- the calculated pKa is preferably 3.5 or more, more preferably 4.5 or more, 5.0 or more, 5.4 or more, 8.0 or more, 8.3 or more.
- the calculated pKa is preferably 10 or less.
- the preferable range of the calculated pKa is preferably 3.5 to 10, 4.5 to 10, 5.0 to 10, 5.4 to 10, 8.0 to 10, or 8.3 to 10.
- the peptide compound or the cyclic peptide compound in the present specification contains an amino acid having a basic side chain that does not intramolecularly hydrogen bond other than an amino acid having a side chain capable of intramolecular hydrogen bonding.
- the Basic pKa of the basic side chain may be 4.0-10.
- the basic side chain has a Basic pKa of preferably 10 or less, more preferably 9.5 or less, 9.0 or less, 8.5 or less, 7.5 or less, or 7.2 or less, and 6.5. The following are particularly preferred.
- the Basic pKa is preferably 4.0 or more.
- the range of the Basic pKa is 4.0 to 10, 4.0 to 9.5, 4.0 to 9.0, 4.0 to 8.5, 4.0 to 7.5, 4.0 to 7.2, 4.0 to 6.5 are preferably exemplified.
- the Basic calculated pKa is preferably 10 or less, 9.5 or less, 9.0 or less, 8.8 or less, 8.6 or less, 8.5 or less, 7.5 or less. Or less, or 6.5 or less is more preferable.
- the Basic calculation pKa is preferably 4.0 or more.
- the range of the basic calculation pKa is 4.0 to 10, 4.0 to 9.5, 4.0 to 9.0, 4.0 to 8.8, 4.0 to 8.6, 4.0. ⁇ 8.5, 4.0 to 7.5, and 4.0 to 6.5 are preferably exemplified.
- the number or ratio of aromatic rings contained in the side chain of the peptide moiety may influence the membrane permeability of the cyclic peptide compound, as shown in the reference example below. If the number or ratio of aromatic rings contained in the side chain of the peptide moiety exceeds a certain number, the probability of obtaining a compound having high membrane permeability may be reduced. That is, the number of aromatic rings can negatively affect the membrane permeability.
- negative impact used in the context of the effect of the invention means an effect that cancels the effect of the invention. For example, assuming that the effect of the invention that should originally be exerted is 100%, and the effect is 30%, 20%, 10%, or 5% or less, there is a "negative effect". be able to.
- the number of aromatic rings contained in the side chain of the peptide moiety of the cyclic peptide compound in the present specification is preferably 5 or less, and 0, 1, 2, for high membrane permeability. , Or 3 is preferred, and 0 to 3, 1 to 3 are preferred ranges.
- the ratio of the number of aromatic rings contained in the side chain of the peptide moiety to the number of amino acids constituting the peptide moiety is preferably 40% or less, 35% or less, 30% or less, 27% or less, 25% or less. , Preferably 20% or less.
- the “number of aromatic rings” refers to the number of aromatic rings contained in the side chain of the peptide portion of the cyclic peptide compound.
- high membrane permeability limits the number of aromatic rings that can be included in the cyclic peptide compounds herein, while including side chains that can form intramolecular hydrogen bonds.
- the amino acid is preferably present in the cyclic portion, which is a site that can contribute to binding with the target molecule. That is, in one embodiment, the peptide compound in the present specification is a cyclic peptide compound, and the cyclic portion of the cyclic peptide compound contains an amino acid containing an aromatic ring in a side chain capable of forming an intramolecular hydrogen bond.
- the number of aromatic rings contained in the cyclic portion of the peptide compound is 1, 2, or 3, and the number of aromatic rings is, for example, 1 to 3 or 2 to 3.
- the peptide compound in the present specification is a cyclic peptide compound, and the cyclic portion of the cyclic peptide compound contains an amino acid containing an aromatic ring in a side chain capable of forming an intramolecular hydrogen bond
- the ratio of the number of aromatic rings contained in the cyclic portion to the total number of aromatic rings contained in the compound is, for example, 30% or more, 40% or more, 60% or more, 80% or more, or 100%.
- the membrane permeability of the peptide compound in the present invention can be measured by the method for measuring membrane permeability described in International Publication No. 2018/124162.
- P app of the peptide compound of the present invention is 1.0 ⁇ 10 ⁇ 7 cm/sec or more, 5.0 ⁇ 10 ⁇ 7 cm/sec or more, 8.0 ⁇ 10 ⁇ 7 cm/sec. Seconds or more is preferable, 9.0 ⁇ 10 ⁇ 7 cm/sec or more is more preferable, 1.0 ⁇ 10 ⁇ 6 cm/sec or more is further preferable, and 3.0 ⁇ 10 ⁇ 6 cm/sec or more is particularly preferable.
- the “membrane permeability coefficient (P app )” in the present specification means a value measured using the measuring method (improved method) described in WO2018/124162.
- ClogP of the cyclic peptide compound in the present specification is preferably 4 or more, more preferably 5 or more, further preferably 6 or more, particularly preferably 8 or more, and 18 or less, 17 or less, 16 or less. It is preferably exemplified, and 4 to 18, 5 to 17, and 6 to 16 are exemplified.
- ClogP in this specification is a distribution coefficient calculated by a computer, and can be calculated using Daylight Version 4.9 of Daylight Chemical Information Systems, Inc.
- the lower limit of ClogP/totalaa of the cyclic peptide compound in the present specification is preferably 1.0 or higher, more preferably 1.1 or higher, and even more preferably 1.2 or higher.
- the upper limit of ClogP/totalaa is preferably 1.8 or less, 1.7 or less, 1.6 or less, 1.5 or less. Examples of the ClogP/totalaa range are 1.0 to 1.8, 1.0 to 1.7, 1.1 to 1.6, and 1.1 to 1.5.
- total aa (also referred to as “total AA”) means the number of amino acids constituting the peptide site of the peptide compound.
- the total aa of a cyclic peptide compound in which the cyclic portion is composed of 10 amino acids and the linear portion is composed of 1 amino acid is 11.
- ClogP/totalaa is calculated by dividing ClogP by totalaa.
- the peptide compounds herein preferably have good metabolic stability.
- the number of amino acids contained in the peptide compound is preferably 8 or more, more preferably 9 or more, still more preferably 11 or more.
- the peptide compound of the present invention has at least one amino acid capable of forming a hydrogen bond (ie, intramolecular hydrogen bond) in its side chain, specifically, for example, one or two amino acids. Includes three, four, or more.
- the side chain of an amino acid in the present specification includes a chain bonded to a carbon atom (eg, ⁇ , ⁇ , or ⁇ -carbon atom) contained in an amino acid and a chain bonded to a nitrogen atom.
- a carbon atom eg, ⁇ , ⁇ , or ⁇ -carbon atom
- the length of the amino acid side chain can be determined by the method described in Reference Example 1.
- the N-terminal of the amino acid unit is capped with an acetyl group and the C-terminal with a methylamino group, and conformation is generated by LowModeMD of the molecular modeling software MOE (Chemical Computing Group), and the side chain site is bound to the atom (
- MOE Chemical Computing Group
- the side chain site is bound to the atom (
- it can be determined by measuring the distance from the ⁇ -carbon atom (C ⁇ carbon)) to the farthest atom on the same side chain (excluding hydrogen atom).
- the “long side chain” refers to a side chain having a length of 5.4 ⁇ or more.
- the side chains of amino acids are preferably long side chains due to intramolecular hydrogen bonding.
- the length of the long side chain is preferably 5.4 angstroms or more, more preferably 5.6 angstroms or more, further preferably 5.8 angstroms or more, and particularly preferably 6.0 angstroms or more.
- the upper limit of the length of the side chain is not particularly limited, but 20 angstroms or less, 15 angstroms or less, 13 angstroms or less, 12 angstroms or less, 11 angstroms or less, 10 angstroms or less, 9.0 angstroms or less, 8.8 angstroms or less, Examples are 8.5 angstroms or less, and 8.0 angstroms or less.
- Examples of the length of the long side chain include 5.4 to 20 angstrom, 6.0 to 20 angstrom, 6.0 to 15 angstrom, 6.0 to 13 angstrom, and 6.0 to 10 angstrom. ..
- the "long side chain” is preferably (i) a side chain that does not contain an amide bond on its side chain or contains one amide bond, and (ii) an amide bond on that side chain. It may be a side chain containing no bond. That is, in one non-limiting embodiment, a cyclic peptide compound having in its cyclic portion an amino acid which does not have two or more amide bonds in its side chain and has a long side chain of 6.0 to 11 angstroms in length. Are preferably exemplified. The long side chain in the present specification may or may not have an aromatic ring.
- the amino acid capable of forming an intramolecular hydrogen bond is preferably a long side chain amino acid.
- the presence or absence of an intramolecular hydrogen bond can be determined by, for example, X-ray structure analysis or chemical shift by 1 H NMR. In general, it is said that when the temperature-dependent chemical shift change amount is ⁇ 2 ppb/K, there is a clear hydrogen bond, and when it is >4 ppb/K, there is no hydrogen bond.
- the conformational analysis by computational chemistry as exemplified in Example 3 is also a method used to judge whether there is a possibility of forming an intramolecular hydrogen bond.
- the peptide compound of the present invention has at least one amino acid capable of forming a pseudo W-membered ring in its side chain, specifically, for example, 1, 2, 3, 4, or Including more.
- the “pseudo W-membered ring” does not have a cyclic structure formed by a covalent bond, but means that a conformation is fixed by an intramolecular hydrogen bond to form a pseudo cyclic structure.
- “W” represents the size of a ring selected from a natural number of 3 or more, and the atom forming the pseudo ring portion is a hydrogen atom that is a proton donor and an atom that is directly bonded to the hydrogen atom, It must contain 3 atoms of proton acceptor capable of forming hydrogen bond with proton donor.
- W representing the ring size of the ring structure of the pseudo ring structure may be 3 or more, preferably 4 to 7, or 5 to 7, and more preferably 5 to 6.
- a hydrogen atom of a hydroxyl group which is a proton donor and an oxygen atom of a hydroxyl group to which the hydrogen atom is directly bonded, and an oxygen atom of an ether group which is a proton acceptor are respectively It corresponds to the above 3 atoms and forms a pseudo 6-membered ring.
- the intramolecular hydrogen bond is characterized in that it can be structurally converted into a cyclized form and a non-cyclized form according to the surrounding environment. By adopting such a cyclized structure, it is expected that a compound having a functional group capable of intramolecular hydrogen bonding is improved in membrane permeability.
- the compound having a functional group for intramolecular hydrogen bonding has a non-cyclized structure, thereby providing a proton donor Both or one of the proton acceptors is expected to bind to the target molecule, and it becomes possible to more efficiently form an intermolecular hydrogen bond between the compound and the target molecule.
- a schematic diagram in which a part of the structure capable of intramolecular hydrogen bonding forms an intermolecular mutual bond with the amide site of the target molecule is shown below (* indicates the ⁇ carbon of Ser(nPrOH) , And ** and *** represent the binding points of the target molecule to the protein).
- an amino acid having a side chain capable of intramolecular hydrogen bonding may form a pseudo 4- to 7-membered ring in the partial structure forming the intramolecular hydrogen bond, and a pseudo 5- to 7-membered ring.
- a pseudo 4- to 7-membered ring in the partial structure forming the intramolecular hydrogen bond
- a pseudo 5- to 7-membered ring are preferably formed, and it is particularly preferable to form a pseudo 5- to 6-membered ring.
- the structure of the present invention that can form an intramolecular hydrogen bond is exemplified below (* represents a point of attachment to Q 1 of Formula 1 or Q 2 of Formula 2).
- amino acids that can form hydrogen bonds within the side chain can be represented by Formula A below.
- R 1 is hydrogen, or C 1 -C 6 alkyl, or a group represented by formula 1 or formula 2, wherein (a) R 2A and or R 2B, either R 2A is hydrogen or C 1 -C 6 alkyl, or a group represented by formula 1 or formula 2, and R 2B is hydrogen or C 1 -C 6 alkyl Alternatively, (b) R 2A and R 2B together with the carbon atom to which they are attached form a 4-6 membered ring. When R 1 is C 1 -C 6 alkyl, R 1 is preferably methyl. When R 2B is C 1 -C 6 alkyl, R 2B is preferably methyl.
- R 2A is preferably a group represented by formula 1 or formula 2 or C 1 -C 6 alkyl.
- R 2A and R 2B together with the carbon atom to which they are bonded form a 4- to 6-membered ring the 4- to 6-membered ring may be cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl. Is preferred.
- R 1 together with the nitrogen atom to which R 1 is bonded, R 2A , and the carbon atom to which R 2A is bonded form a 4- to 6-membered heterocycle
- R 2B is hydrogen or C 1 -C 6 alkyl.
- R 2B is preferably methyl.
- the 4- to 6-membered heterocycle may have one or more substituents selected from the group consisting of the group represented by Formula 1 or Formula 2, —OH, and an alkoxy group.
- the 4- to 6-membered heterocycle is preferably pyrrolidine (that is, a proline-like structure).
- R 1 , R 2A and R 2B when R 2A is a group represented by formula 1 or formula 2, R 1 is hydrogen or C 1 -C 6 alkyl, and R 2B is hydrogen or is preferably a C 1 -C 6 alkyl, it is preferred that the C 1 -C 6 alkyl is methyl.
- R 1 is a group represented by formula 1 or formula 2
- R 2A is hydrogen or C 1 -C 6 alkyl
- R 2B is hydrogen, C 1 -C 6 alkyl, or R 2A and R 2B are preferably taken together to form a 4 to 6 membered ring, and the C 1 -C 6 alkyl is preferably methyl.
- R 3 is a single bond or —CHR 4 —, wherein R 4 is hydrogen, C 1 -C 4 alkyl, or is represented by formula 1 or formula 2. It is a base.
- R 4 is preferably hydrogen or a group represented by Formula 1 or Formula 2.
- R 1 is a group represented by formula 1 or formula 2
- R 2A is hydrogen, or C 1 -C 6 alkyl, preferably hydrogen
- R 2B is hydrogen or C 1 -C 6 Alkyl, preferably hydrogen
- R 3 is a single bond, or —CHR 4 —
- R 4 is hydrogen, or C 1 -C 4 alkyl, preferably hydrogen.
- R 1 is hydrogen or C 1 -C 6 alkyl
- R 2A is a group represented by formula 1 or formula 2
- R 2B is hydrogen or C 1 -C 6 alkyl, preferably hydrogen
- R 3 is a single bond, or —CHR 4 —, preferably a single bond
- R 4 is hydrogen or C 1 -C 4 alkyl, preferably hydrogen.
- R 1 is hydrogen or C 1 -C 6 alkyl
- R 2A is hydrogen, or C 1 -C 6 alkyl, preferably hydrogen
- R 2B is hydrogen, or C 1 -C 6 alkyl, preferably Is hydrogen
- R 3 is —CHR 4 —
- R 4 is a group represented by Formula 1 or Formula 2.
- R 1 a nitrogen atom to which R 1 is bound, R 2A , and a carbon atom to which R 2A is bound together form a 4- to 6-membered heterocycle (preferably a pyrrolidine ring),
- the ring has a group represented by formula 1 or formula 2,
- R 2B is hydrogen or C 1 -C 6 alkyl, preferably hydrogen
- R 3 is a single bond, or —CHR 4 —, preferably It is a single bond
- R 4 is hydrogen or C 1 -C 4 alkyl, preferably hydrogen.
- Formula 1 and Formula 2 are respectively the following formulas: Is shown in the formula, and * in the formula represents a point of attachment.
- the amino acid represented by formula A contains at least one of the groups represented by formula 1 or formula 2.
- the amino acid represented by Formula A can include either one of the groups represented by Formula 1 or Formula 2, in which case any of R 1 , R 2 , or R 4 in Formula A can be included.
- Q 1 is a single bond, C 1 -C 4 alkylene, or C 2 -C 4 heteroalkylene containing one oxygen atom.
- Q 1 is preferably —CH 2 — or —(CH 2 ) 2 —.
- a 1 is —O— or —S—, preferably —O—.
- L 1 is substituted with one or more substituents selected from the group consisting of fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O). It may be a straight chain C 1 -C 3 alkylene.
- L 1 examples include —CH 2 —, —(CH 2 ) 2 —, —(CH 2 ) 3 —, —CH 2 CH(CH 3 )—, —CH 2 CH(CF 3 )—, — CH 2 C(CH 3 ) 2 —, —(CH 2 ) 2 CH(CH 3 )—, —CH 2 C(CH 3 ) 2 CH 2 —, —(CH 2 ) 2 C(CH 3 ) 2 —, —CH 2 CO— and the like can be mentioned. In one aspect, it is preferred that L 1 is not unsubstituted —(CH 2 ) 2 —.
- X is —OH, —NR Z1 R Z2 , or —CONR Z1 R Z2 , or contains 1 to 3 heteroatoms, and is substituted with oxo or one or more halogens. 5 to 6 membered saturated or unsaturated heterocyclyl, wherein R Z1 and R Z2 are independently hydrogen, —OH, C 1 -C 4 alkyl, and C 1 -C 4 alkylsulfonyl. Is selected from the group consisting of When X is -NR Z1 R Z2 , X is preferably -NH 2 , -NHMe, -NHEt, -NH(nPr), -NH(iPr), -NH(tBu).
- X is preferably -CONH 2 , -CONHMe, -CONHEt, -CONH(nPr), -CONH(iPr), -CONH(tBu).
- X is oxo or a 5- to 6-membered saturated or unsaturated heterocyclyl optionally substituted with one or more halogen
- the saturated or unsaturated heterocyclyl is 4,5-dihydro-1,2,4 -Oxadiazolyl, 4,5-dihydro-1,2,4-thiadiazolyl and pyrrolidyl are preferable, and the following groups are represented as X: Is preferred.
- Q 1 is C 1 -C 4 alkylene, preferably —CH 2 —
- a 1 is —O— or —S—, preferably —O—
- L 1 is C 1 -C
- X is --OH.
- Q 1 is C 1 -C 4 alkylene, preferably —CH 2 —
- a 1 is —O— or —S—, preferably —O—
- L 1 is C 1 -C. 2 alkyl groups, C 1 -C 2 fluoroalkyl, and straight chain C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of oxo ( ⁇ O), preferably Is —CH 2 —
- X is —NR Z1 R Z2
- R Z1 is C 1 -C 4 alkyl
- R Z2 is hydrogen.
- Q 1 is C 1 -C 4 alkylene, preferably —CH 2 —
- a 1 is —O— or —S—, preferably —O—
- L 1 is C 1 -C. 2 alkyl groups, C 1 -C 2 fluoroalkyl, and straight chain C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of oxo ( ⁇ O), preferably Is —CH 2 —
- X is —CONR Z1 R Z2
- R Z1 is C 1 -C 4 alkyl and R Z2 is hydrogen.
- Q 1 is C 1 -C 4 alkylene, preferably —CH 2 —
- a 1 is —O— or —S—, preferably —O—
- L 1 is C 1 -C. 2 alkyl groups, C 1 -C 2 fluoroalkyl, and straight chain C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of oxo ( ⁇ O), preferably Is —CH 2 —, wherein X contains 1 to 3 heteroatoms and is 5 to 6 membered saturated or unsaturated optionally substituted by oxo or one or more halogen (preferably fluorine). It is a saturated heterocyclyl.
- Q 1 is a single bond
- a 1 is —O— or —S—, preferably —O—
- L 1 is a C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, And a linear C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of and oxo ( ⁇ O), and X is —OH.
- Q 2 is a single bond, C 1 -C 4 alkylene, or C 2 -C 4 heteroalkylene containing one oxygen atom.
- Q 2 is preferably —CH 2 — or —(CH 2 ) 2 —.
- L 2 is a single bond, or one or more selected from the group consisting of fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O) Is a linear C 1 -C 3 alkylene which may be substituted by the substituent of.
- Specific examples of L 2 include a single bond, —CH 2 —, —CO—, —(CH 2 ) 2 —, —(CH 2 ) 3 —, —CH 2 CH(CH 3 )—, —CH 2 CH.
- Y is —OH, C 1 -C 4 alkylsulfonylamino, —NR Z3 R Z4 , or —CONR Z3 R Z4 , or contains 1 to 3 heteroatoms, and is oxo or halogen. 5 to 6 membered saturated or unsaturated heterocyclyl optionally substituted by wherein R Z3 and R Z4 are independently hydrogen, —OH, C 1 -C 4 alkyl, and C 1 -C It is selected from the group consisting of 4 alkylsulfonyl.
- Y is preferably —NH 2 , NHMe, NHEt, NH(nPr), NH(iPr), NH(tBu).
- Y is C 1 -C 4 alkylsulfonylamino
- Y is preferably methylsulfonylamino.
- L 2 is —CO—.
- a 2 is a single bond, —O—, or —S—, and preferably —O—.
- Z is hydrogen or C 1 -C 4 alkyl, and when Z is C 1 -C 4 alkyl, the C 1 -C 4 alkyl is preferably methyl.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —
- L 2 is fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo
- Y is —NR Z3 R Z4 , preferably methyl.
- a 2 is —O—, or —S—, preferably —O—, and Z is C 1 -C 4 alkyl, preferably methyl.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —, L 2 is a single bond, and Y is —CONR Z3 R Z4 , preferably methylaminocarbonyl or —CON(OH).
- a 2 is —O—, or —S—, preferably —O—, and Z is C 1 -C 4 alkyl, preferably methyl.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —
- L 2 is fluorine, a C 1 -C 2 alkyl group, a C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O) Is a straight chain C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of, preferably —CO—
- Y is —NR Z3 R Z4 , preferably — N(OH)Me
- a 2 is a single bond
- Z is hydrogen.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —, L 2 is a single bond, Y is —CONR Z3 R Z4 , preferably —CON(OH)Me, A 2 is a single bond and Z is hydrogen.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —
- L 2 is fluorine, C 1 -C 2 alkyl group, C 1 -C 2 fluoroalkyl, and oxo ( ⁇ O)
- a linear C 1 -C 3 alkylene optionally substituted by one or more substituents selected from the group consisting of, preferably —CO—
- Y is a C 1 -C 4 alkylsulfonylamino, It is preferably methylsulfonylamino
- a 2 is —O—, or —S—, preferably —O—
- Z is C 1 -C 4 alkyl, preferably methyl.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —
- L 2 is a single bond
- Y is C 1 -C 4 alkylsulfonylaminocarbonyl, preferably methylsulfonylaminocarbonyl.
- a 2 is —O—, or —S—, preferably —O—
- Z is C 1 -C 4 alkyl, preferably methyl.
- Q 2 is C 1 -C 4 alkylene, preferably —CH 2 —, L 2 is a single bond, Y is —OH, A 2 is —O—, or —S—, It is preferably —O— and Z is C 1 -C 4 alkyl, preferably methyl.
- amino acid capable of forming a hydrogen bond in the side chain and/or the amino acid capable of forming a pseudo 4- to 7-membered ring in the side chain include the following amino acids.
- the amino acid represented by the formula A of the present invention when the amino acid represented by the formula A of the present invention is contained in a peptide compound, it is preferable to link the adjacent amino acid by an amino group and a carboxyl group of the main chain of the amino acid.
- the amino acid represented by the formula (A) can also be represented as follows (* represents a bonding point with an adjacent amino acid in the peptide compound).
- the present invention relates to a protected amino acid in which an amino group and/or a carboxyl group of the amino acid capable of forming an intramolecular hydrogen bond are protected with a protecting group.
- the protecting group for the amino group of the protected amino acid include Fmoc group, Boc group, Cbz group, Alloc group, nosyl group, dinitronosyl group, t-Bu group, trityl group and cumyl group.
- the protective group for the carboxyl group of the protected amino acid include a methyl group, an allyl group, a t-Bu group, a trityl group, a cumyl group, a methoxytrityl group and a benzyl group.
- these protecting groups can be introduced using the method described in "Greene's, "Protective Groups in Organic Synthesis” (5th edition, John Wiley & Sons 2014).
- the position of the proton donor is limited to the position close to the main chain amide bond that can form hydrogen bonds. It will be what was done.
- the amino acid of the present specification can form a hydrogen bond “inside the side chain” thereof, which may increase the degree of freedom in side chain design. For example, it is possible to lengthen the side chains while maintaining the formation of hydrogen bonds that may mask the protons, which may increase the likelihood that the protons on the side chains will be used for interaction with the target.
- an amino acid having an ether bond in its side chain can be produced by the following two synthetic methods.
- Manufacturing method 1 A compound having an aziridine in which an amino group and a carboxylic acid group are protected is reacted with an alcohol having an appropriate corresponding protecting group in the presence of Lewis acid to perform a ring-opening reaction, and then an unnecessary protecting group is deprotected.
- Cbz group as P 1 Fmoc group, Alloc group, and nosyl groups (Ns group), as the P 2 Me group, an allyl group, can be used a benzyl group.
- the Lewis acid used in the ring-opening reaction is preferably exemplified by a catalytic amount of BF 3 .OEt 2 .
- Dichloromethane is preferably exemplified as the solvent, but alcohol (ROH) used as a reagent can also be used as the solvent.
- R contains a functional group that may participate in the reaction, it is preferable to protect the functional group with a protective group that is stable in the presence of Lewis acid.
- R contains a hydroxyl group, it is preferable to protect the hydroxyl group with a benzyl group or the like, and when R contains a secondary amino group, the amino group is protected with an Alloc group or the like.
- the desired amino acid (P 3 is a Cbz group, Fmoc group, Boc group, Alloc group, nosyl group (Ns group), etc.) Protected amino acids or unprotected amino acids where P 3 is hydrogen).
- Manufacturing method 2 After performing an alkylation reaction between an amino group-protected amino acid having a hydroxyl group in the side chain and an ⁇ -halocarbonyl compound such as 2-bromo-N-methylacetamide, or bromoacetonitrile, the following functional group conversion is performed.
- the production shown in the scheme can be carried out.
- P 1 is preferably a protective group that is stable under basic conditions such as a Cbz group, a Boc group, and a Trt group.
- Preferable examples of the ⁇ -halocarbonyl compound include those in which R a is NMe, NtBu, or the like.
- X 1 is halogen and is preferably a bromo group.
- Examples of the base used in the alkylation reaction include NaH, NaOtBu, and NaOt-Pent, and the hydroxyl group of the amino acid is selectively selected by using 2 equivalents or more of the amino acid having a hydroxyl group in the side chain and having an amino group protected.
- the site can be alkylated.
- Preferable examples of the solvent used include DMF and DMI.
- an amino acid basic skeleton can be constructed by constructing a heterocycle based on the obtained nitrile group. In the scheme below, hydroxylamine is allowed to act on the resulting nitrile group, and then an oxadiazolone ring cyclized with CDI is constructed.
- Desired amino acid is Cbz group by performing the appropriate transformation of the protective group after building the backbone of an amino acid, Fmoc group, Boc group, Alloc group, nosyl group (Ns group) protected amino acids, etc. Alternatively, it can be converted to an unprotected amino acid) in which P 3 is hydrogen.
- the phenylalanine derivative is a method described in Jackson-Negishi coupling (Journal of Organic Chemistry, 2010, 75, 245) or asymmetric benzylation of benzophenone imine of glycine tert-butyl ester using a phase transfer catalyst (J. Am. Chem. Soc. 1999, 121, 6519) and the like. It can also be produced by carrying out a cross-coupling reaction involving decarboxylation using an N-hydroxyphthalimide ester (NHPI ester) represented by the following general formula and an aromatic iodine compound or an aromatic bromo compound as raw materials. ..
- NHPI ester N-hydroxyphthalimide ester
- P 1 a group that is stable under cross-coupling conditions is used, and preferably a Fmoc group is exemplified.
- R 4 -X 1 is an aromatic halogen compound, and R 4 represents an aromatic moiety.
- R 4 contains a functional group that can be converted under the cross-coupling reaction conditions, it is preferably protected with a protective group that is stable under the cross-coupling reaction conditions.
- R 4 contains a hydroxyl group, it is preferable to protect the hydroxyl group with a tetrahydropyranyl (THP) group or the like.
- THP tetrahydropyranyl
- X 1 is a halogen, preferably a bromo (Br) group or an iodo (I) group.
- the catalyst used in the cross-coupling reaction is preferably nickel and zinc, and the ligand of the catalyst is 4,4′-di-tert-butyl-2,2′-bipyridyl (dtbbby). Is preferred. It is preferable to use dimethylacetamide as the solvent.
- a desired amino acid (P 3 is a Cbz group, an Fmoc group, a Boc group, an Alloc group, a nosyl group (P 3 is Ns group) or the like, or an unprotected amino acid in which P 3 is hydrogen.
- P 3 is a side chain site on the N atom of the amino acid.
- the corresponding ⁇ -amino acid can be produced by using Arndt-Eistert synthesis (organic synthetic strategy learned from personal name reaction, p18).
- a ring-opening reaction with ammonia is performed on an epoxide having a corresponding functional group, a protecting group is introduced at the amine site of the resulting compound, and then the resulting secondary alcohol is converted to a leaving group. It can be produced by converting to a nitrile group accompanied by steric inversion by a nucleophilic substitution reaction, and then converting to a carboxylic acid.
- R contains a functional group involved in the reaction, it is preferable to protect with a suitable protecting group.
- R contains a hydroxyl group, it is preferable to protect the hydroxyl group with a Trt group.
- Preferred examples of P 1 which is the amino-protecting group obtained after the ring-opening reaction include Cbz group, Boc group and Alloc group.
- the produced secondary alcohol is converted into a leaving group X 2 (preferably, X 2 is exemplified by a mesyl group (OMs group) and the like).
- a basic skeleton of an amino acid can be constructed by converting the resulting leaving group into a nitrile group by nucleophilic substitution reaction with cyanohydrin (CN ⁇ ) with steric inversion and converting it into a carboxylic acid.
- the conversion of a nitrile group into a carboxylic acid is exemplified by hydrolysis under acidic conditions.
- the desired amino acid (P 3 was protected with a Cbz group, Fmoc group, Boc group, Alloc group, nosyl group (Ns group), etc. Amino acid or an unprotected amino acid in which P 3 is hydrogen).
- a cyclic amino acid having a hydroxyl group and protected with an amino group and a halogen compound are subjected to an alkylation reaction using a base, and then a suitable functional group is converted to give a cyclic amino acid as shown in the following scheme.
- the cyclic amino acid used as a starting material is preferably a 4- to 8-membered cyclic amino acid, and a 5-membered ring is exemplified in the following scheme.
- the position of the hydroxyl group present in the cyclic portion is not particularly limited, but as shown in the general formula, it is preferably at a position separated by 2 or more atoms from the N atom of the amino acid.
- Cbz group, Boc group, Alloc group, nosyl group (Ns group) or the like can be used as P 1 .
- the base used in the alkylation reaction is preferably sodium hydride (NaH), and the solvent is preferably dimethylformamide.
- the halogen compound (R—X) when the halogen compound (R—X) has a functional group on R that may participate in the reaction, it is preferable to protect the functional group with a stable protecting group under basic conditions. ..
- R when R contains a hydroxyl group, it is preferable to protect the hydroxyl group with a silyl ether such as a tetrahydropyranyl (THP) group, a benzyl (Bn) group and a t-butyldimethylsilyl (TBDMS) group.
- a silyl ether such as a tetrahydropyranyl (THP) group, a benzyl (Bn) group and a t-butyldimethylsilyl (TBDMS) group.
- THP tetrahydropyranyl
- Bn benzyl
- TDMS t-butyldimethylsilyl
- X 1 is halogen, and examples thereof include a chloro (Cl) group, a bromo (Br) group, and an iodo (I) group.
- the desired amino acid (P 3 is a Cbz group, Fmoc group, Boc group, Alloc group, nosyl group (Ns group), etc.) Protected amino acids or unprotected amino acids where P 3 is hydrogen).
- Process 7 According to the scheme below, the above amino acid undergoes a cyclization reaction using paraformaldehyde under acidic conditions, and then the resulting cyclized product is subjected to ring opening under acidic conditions to reduce the resulting imine. , N-methylation can be performed.
- the basic unit is not particularly limited as long as it is a combination having an Fmoc-protected amino group and a carboxyl group.
- a dipeptide may be used as a basic unit.
- the basic unit placed at the N-terminus may be other than the Fmoc amino acid.
- it may be a Boc amino acid or a carboxylic acid analog having no amino group.
- a side chain carboxyl group of an amino acid having a carboxyl group on the main chain carboxyl group or a side chain whose main chain carboxyl group is protected by a suitable protecting group is supported on a solid phase by a chemical reaction with a functional group of a solid phase carrier.
- the Fmoc group is deprotected with a base such as piperidine or DBU, and the newly formed amino group is subjected to a condensation reaction with a protected amino acid having a basic unit carboxyl group to be subsequently added to form a peptide bond.
- a base such as piperidine or DBU
- various combinations such as a combination of DIC and HOBt, a combination of DIC and HOAt, and a combination of HATU and DIPEA can be used as the carboxyl group activator.
- the desired peptide sequence can be generated by repeating the Fmoc-eliminating group and the subsequent peptide bond formation reaction.
- cleavage from the solid phase and, if necessary, deprotection of the protecting group of the introduced side chain functional group are performed. It is also possible to carry out structural conversion or cyclization of the peptide before excising it from the solid phase. Cleavage from the solid phase and deprotection may be performed under the same conditions, for example, 90:10 TFA/H 2 O, or may be deprotected under different conditions if necessary.
- Cleavage from the solid phase some can be cleaved with a weak acid such as 1% TFA, and Pd or the like can be used as a protecting group to utilize the orthogonality of both chemical reactions. Steps such as cyclization can also be carried out between or at the end of these steps.
- a side chain carboxylic acid can be condensed with an amino group of the N-terminal main chain, or a side chain amino group can be condensed with a carboxylic acid of a C-terminal main chain.
- the reaction orthogonality is required between the carboxylic acid on the C-terminal side and the side-chain carboxylic acid to be cyclized, or between the main chain amino group or hydroxy group on the N-terminal side and the side-chain amino group to be cyclized.
- the protecting group is selected in consideration of the orthogonality of the protecting group.
- the reaction product thus obtained can be purified by a reverse phase column, a molecular sieve column or the like.
- the resin for solid phase synthesis a commercially available resin can be used, and examples thereof include CTC resin, Wang resin, and SASRIN resin.
- the compound when the defined group undergoes an undesired chemical conversion under the conditions of the method for carrying out the method, for example, by protecting the functional group, deprotecting it,
- the compound can be prepared.
- the protective group selection and desorption operations include the method described in “Greene's, “Protective Groups in Organic Synthesis” (5th edition, John Wiley & Sons 2014)”, and these are used as reaction conditions. It may be used as appropriate. Further, the order of reaction steps such as introduction of substituents can be changed as necessary.
- the protecting group for the amino group is exemplified by Fmoc, Boc, Cbz, or Alloc group.
- carbamate groups can be introduced by reacting an amino group with a carbamate agent in the presence of a base catalyst.
- a carbamate agent include Boc 2 O, BocOPh, FmocOSu, FmocCl, CbzCl, AllocCl and the like.
- the base catalyst include lithium carbonate, sodium carbonate, sodium hydrogen carbonate, potassium carbonate, potassium hydrogen carbonate, cesium carbonate, cesium hydrogen carbonate, lithium hydroxide, sodium hydroxide, potassium hydroxide, cesium hydroxide, sodium phosphate and phosphorus.
- Examples thereof include potassium acid salt, N-methylmorpholine, triethylamine, diisopropylethylamine, N,N-dimethylaminopyridine and the like.
- the carbamate group which is a protective group for the amino group, can be removed under basic conditions, acidic conditions, hydrogenolysis reaction conditions, or the like.
- Method for Translational Synthesis of Peptide Compound As a method for translational synthesis of a peptide compound in the present specification, a method of synthesizing using a cell-free translation system is exemplified, and the method includes a method of synthesizing by a reconstituted cell-free translation system. It is preferable to use the reconstituted cell-free translation system because the factor to be eliminated can be removed.
- the present specification provides a method for translationally synthesizing a peptide compound according to the present specification, which comprises at least one kind, two or more kinds, or three or more kinds of amino acids in the present specification.
- Such translation synthesis methods may include, but are not limited to, the following (i) and (ii): (i) a step of preparing a tRNA having at least one kind, two or more kinds, or three or more kinds of amino acids in the present specification bound thereto; (ii) a step of translating a nucleic acid containing at least one codon corresponding to the anticodon of the tRNA in a cell-free translation system to obtain the peptide compound.
- the method of translational synthesis herein may be a method of translational synthesis of a cyclic peptide compound.
- the "cell-free translation system” is a combination of a ribosome extracted from a cell, a protein factor group involved in translation, an energy source such as tRNA, amino acid, ATP, and its regeneration system. However, it is not limited as long as it can translate mRNA into protein. A system in which a translation synthesis reaction is in progress may also be included in the "cell-free translation system" in the present specification.
- the cell-free translation system in the present specification may include a nucleic acid serving as a template for translating a peptide, and may further include an initiation factor, an elongation factor, a dissociation factor, an aminoacyl-tRNA synthetase, and the like. .
- the cells for purifying the factor can include, for example, prokaryotic cells or eukaryotic cells.
- Prokaryotic cells can include E. coli cells, hyperthermophilic cells, or Bacillus subtilis cells.
- Known eukaryotic cells are yeast cells, wheat germ, rabbit reticulocytes, plant cells, insect cells, or animal cells.
- tRNAs and aminoacyl-tRNA synthetases ARS
- artificial tRNAs and artificial aminoacyl-tRNA synthetases that recognize unnatural amino acids can be used.
- RNA polymerase such as T7 RNA polymerase can be added to the cell-free translation system to cause transcription from the template DNA.
- the phrase "comprising a substance having a cell-free translation system” means that the substance is synthesized and contained in the system during the translation synthesis process even if the substance is not contained at the initiation of translation synthesis. It also includes a mode that becomes. For example, when a tRNA in which an amino acid is acylated is synthesized in the process of translation synthesis, it is understood that the cell-free translation system includes the aminoacyl tRNA.
- PURESYSTEM (registered trademark) (BioComber, Japan) is a reconstituted cell-free translation that extracts and purifies protein factors, energy regeneration system enzymes, and ribosomes required for E. coli translation, and mixes them with tRNA, amino acids, ATP, GTP, etc. It is a system. Not only is the content of impurities small, but since it is a reconstituted system, it is possible to easily prepare a system that does not contain the protein factor or amino acid to be excluded. ((i)Nat Biotechnol. 2001;19:751-5. Cell-free translationreconstituted with purified components.
- translational synthesis includes, for example, protein factors necessary for translation of E. coli (methionyl tRNA transformylase, EF-G, RF1, RF2, RF3, RRF, IF1, IF2, IF3, EF-Tu, EF-Ts, ARS (AlaRS, ArgRS, AsnRS, AspRS, CysRS, GlnRS, GluRS, GlyRS, HisRS, IleRS, LeuRS, LysRS, MetRS, PheRS, ProRS, SerRS, ThrRS, TrpRS, TyrRS, ValRS )), ribosome, amino acid, creatine kinase, myokinase, inorganic pyrophosphatase, nucleoside diphosphate kinase, E.
- E. coli methionyl tRNA transformylase, EF-G, RF1, RF2, RF3, RRF, IF1, IF2, IF3, EF-Tu, EF-Ts
- coli tRNA creatine phosphate, potassium glutamate, HEPES-KOH pH 7.6, magnesium acetate, spermidine, dithiothrey. It can be performed by adding mRNA to a known cell-free translation system such as PURE SYSTEM in which Tol, GTP, ATP, CTP, UTP and the like are appropriately selected and mixed.
- PURE SYSTEM in which Tol, GTP, ATP, CTP, UTP and the like are appropriately selected and mixed.
- T7 RNA polymerase is added, transcription and translation from the template DNA containing the T7 promoter can be performed in a coupled manner.
- a peptide compound containing an unnatural amino acid can be translationally synthesized by adding to the system an unnatural amino acid group (for example, F-Tyr) allowed by a desired aminoacyl-tRNA group or aminoacyl-tRNA synthetase (ARS) ( Kawakami T, et al. Ribosomal synthesis polypeptoids and peptoid-peptide hybrids. J Am Chem Soc. 2008, 130, 16861-3., Kawakami T, et al. Diverse backbone-cyclized peptides via codon at , 5, 888-90.).
- an unnatural amino acid group for example, F-Tyr
- ARS aminoacyl-tRNA synthetase
- a peptide compound containing an unnatural amino acid can be translationally synthesized by including a modified form of ARS in the system instead of or in addition to the natural ARS and by including an unnatural amino acid group in the system.
- it is possible to increase the efficiency of translation introduction of unnatural amino acids by utilizing variants such as ribosomes and EF-Tu (DedkovaLM, et al. Construction of modified ribosomes for incorporation D-amino acids toto proteins. Biochemistry. 2006, 45, 15541-51., Doi Y, et al. Elongation factor Tumutants expand amino acid tolerance of protein biosynthesis system. J AmChem Soc. 2007, 129, 14458-62., Park HS, etal. Expanding the genetic code of Escherichia coli with phosphoserine.Science. 2011, 333, 1151-4.).
- the cell-free translation system herein may be a translation system for producing a peptide compound, preferably a cyclic peptide compound. Is a translation system for manufacturing.
- the translation system herein may include 5 to 32, 5 to 28, or 5 to 20 unnatural amino acids, 8 to 20, 10 to 20, Preferable examples are 13 to 20 kinds. In one embodiment, 50% or more, 60% or more, 70% or more, 80% or more of the number of types of amino acids contained in the translation system in the present specification may be unnatural amino acids.
- the translation system herein may include 5 to 28 kinds, or 5 to 20 kinds of N-substituted amino acids, and 5 to 18 kinds and 5 to 15 kinds are preferably exemplified.
- 40% or more, 50% or more, 60% or more, 70% or more of the number of types of amino acids contained in the translation system in the present specification may be N-substituted amino acids.
- the N-substituted amino acid herein may mean an N-alkyl amino acid or an N-methyl amino acid, but even in that case, it is not necessary for the translation system in the present specification to include other N-substituted amino acids. I do not deny it.
- the translation system in the present specification may be adjusted so that the average number of aromatic rings contained in the peptide compound produced using the translation system is within a certain range. ..
- the average ratio of the number of amino acids having an aromatic ring to the number of amino acids constituting the cyclic portion is 40% or less, 35% or less, 30% or less, 27% or less, 25%. Or less, or 20% or less, or the average number of aromatic rings contained in the side chain of the cyclic part of the peptide compound having a cyclic part composed of 8 to 11 amino acids becomes 0 to 3
- the translation system herein may be adjusted.
- the adjusting method is not particularly limited, and examples thereof include adjusting the number of kinds of amino acids having an aromatic ring in the number of kinds of amino acids contained in the translation system.
- the translation system herein may include an amino acid having an aromatic ring, and the ratio of the number of amino acid types having an aromatic ring to the number of amino acid types included in the translation system is 40% or less is preferable, and 35% or less, 30% or less, 27% or less, 25% or less, 20% or less are preferably exemplified.
- the cell-free translation system herein comprises a cell-free translation system for producing a peptide compound of the present specification, which comprises at least one amino acid of the present specification, two or more, or three or more of them. It may be a translation system.
- Such cell-free translation systems may include, but are not limited to, the following (i) and (ii): (i) tRNA having at least one kind, two or more kinds, or three or more kinds of amino acids in the present specification bound thereto; (ii) a nucleic acid encoding the peptide compound,
- the nucleic acid may include at least one codon corresponding to the anticodon of the tRNA.
- tRNA For translational introduction of unnatural amino acids into peptides, tRNA ((i)Biochemistry. 2003;42:9598-608. Adaptation of an orthogonal archaeal leucyl-tRNA and synthetase pair for four-base, amber, and opal suppression.Anderson JC, Schultz PG., (ii)Chem Biol. 2003;10:1077-84.
- a solid-phase ribozyme aminoacylation system to reprogram the genetic code.Murakami H, Kourouklis D , Suga H.
- the following five methods can be used for aminoacylating tRNA.
- aminoacyl-tRNA synthetase is prepared for each amino acid as an enzyme for aminoacylation of tRNA. Therefore, as the first method, a method of utilizing that certain ARS allows unnatural amino acids such as N-MeHis and a mutant aminoacyl-tRNA synthetase that allows unnatural amino acids are prepared and used. ((I)Proc Natl Acad Sci U S A. 2002;99:9715-20.
- aminoacyl-tRNA can be obtained by ligating the 3'-terminal CCA sequence of tRNA without CA and separately prepared aminoacylated pdCpA with RNA ligase (Biochemistry. 1984;23:1468). -73.T4 RNAligase mediated preparation of of novel "chemically misacylated" tRNAPheS.Heckler TG, Chang LH, Zama Y, Naka T, ChorghadeMS, Hecht SM.). There is also aminoacylation by flexizyme, which is a ribozyme that supports active esters of various unnatural amino acids on tRNA (J Am Chem Soc.
- aminoacylation is also possible by adding to the tRNA the amino acid active ester linked to the PNA complementary to the 3'end of the tRNA (J Am ChemSoc. 2004;126:15984-9. In situ chemical aminoacylation with amino acid thioesters linkedto to a peptide.Nucleic acid.NinomiyaK,MinohataT,NishimuraM,Sisido M.).
- aminoacyl-tRNA can be produced using the following method. Prepare a template DNA that encodes the desired tRNA sequence and has a T7, T3 or SP6 promoter placed upstream, and synthesizes RNA by transcription using RNA polymerase suitable for the promoter such as T7 RNA polymerase or T3, SP6 RNA polymerase. You can do it. It is also possible to extract and purify tRNA from cells, and to extract the target tRNA produced using a probe having a sequence complementary to the tRNA sequence. At this time, a cell transformed with the target tRNA expression vector can also be used as a source. RNA having a target sequence can also be synthesized by chemical synthesis.
- an aminoacyl-tRNA can be obtained by ligating the tRNA obtained by removing the CA from the CCA sequence at the 3′ end thus obtained with separately prepared aminoacylated pdCpA or pCpA with RNA ligase (the pdCpA method). , PCpA method).
- the tRNA is useful in the production of peptide compounds.
- full-length tRNA is prepared, and aminoacylation with flexizyme, which is a ribozyme supporting various active esters of unnatural amino acids on tRNA, is also possible.
- naturally occurring ARS or its variants can also be used to make aminoacyl tRNAs.
- the aminoacyl tRNA once consumed in the translation system can be regenerated by the natural ARS or its modified form. Therefore, it is necessary to make a large amount of aminoacyl tRNA produced in advance in the translation system. Absent. Such ARS variants are described in WO2016/148044. These aminoacyl-tRNA production methods can be combined as appropriate.
- the cyclic portion of the peptide compound is formed by subjecting a linear peptide compound to a cyclization reaction.
- the “cyclization reaction” means a reaction for forming a cyclic part at the peptide site of a peptide compound.
- a peptide compound obtained by further chemically modifying or reacting a peptide compound having a cyclic portion formed by a cyclization reaction is also included in the peptide compound in the present specification. Moreover, you may perform chemical modification etc. before cyclization reaction.
- Scheme A shows an example of the cyclization reaction of the peptide moiety of the peptide compound in the present specification.
- a black circle ( ⁇ ) unit main chain unit
- a D unit aspartic acid unit
- a triangle ( ⁇ ) unit cyclized N-terminal unit
- White circles represent resins for solid phase synthesis.
- the ring portion is a portion composed of one triangular unit, nine black circle units and one D unit. Each unit may be the same amino acid or different amino acids.
- the cyclic peptide of the present invention can be produced, for example, by the method described in WO 2013/100132.
- the “unit” means any of an amino acid after synthesis of the chain peptide compound before cyclization, an amino acid after cyclization, and an amino acid at the time of completion of chemical modification after cyclization.
- the amino acid residue at the time of completion of the chemical modification after the cyclization includes, for example, an amino acid residue chemically or skeleton-converted by the chemical modification after the production of the peptide compound.
- a peptide compound or a library of nucleic acids encoding them in the present disclosure is a library consisting essentially of the peptide compounds described herein or nucleic acids encoding them, respectively.
- the libraries herein include libraries containing the peptide compounds herein, and libraries containing nucleic acids encoding the peptide compounds herein.
- the library in the present specification includes a peptide compound library and a peptide compound-nucleic acid complex library in the present specification. Among them, a cyclic peptide compound or a cyclic peptide-nucleic acid complex library is preferable, and a cyclic peptide-particularly A library of mRNA complexes is preferred.
- a display library is preferable as the library.
- Examples of the display library include a library using a display, and among them, an mRNA display library, a DNA display library, and a ribosome display library are preferable, and an mRNA display library is more preferable.
- the present inventors think as follows. In peptide libraries so far, it has been accidentally relied on to mask the proton donors contained in the side chains of the amino acids that make up the peptide by intramolecular hydrogen bond formation. In particular, in a peptide library in which constituent amino acids are randomly arranged to give diversity, it has been difficult to form hydrogen bonds between amino acids in a targeted manner. In the case of a peptide compound containing an amino acid of the present specification, an amino acid in which a hydrogen bond has already been formed at the proton donor site contained in the side chain of an amino acid is used, and therefore a hydrogen atom in the molecule at the side chain site of the peptide compound The bond can be formed intentionally.
- the peptide compound library obtained by using the amino acid of the present specification is a peptide compound having a high frequency of appearance of a peptide compound capable of masking a side-chain proton donor by forming a hydrogen bond in the molecule, that is, a peptide compound having high membrane permeability. Can be more likely to be included.
- a peptide compound may be a “peptide compound encoded by a nucleic acid”.
- the term "peptide compound encoded by a nucleic acid” is not limited to a peptide compound directly synthesized by translating the nucleic acid as a template, but includes a peptide compound that can be synthesized by post-translational modification. There is.
- the cyclic peptide compound may be referred to as a cyclic peptide compound encoded by the nucleic acid. ..
- a peptide compound that is translationally synthesized using a nucleic acid as a template, or a complex of the peptide compound that has been post-translationally modified with the nucleic acid is also encoded by the nucleic acid. Included in (cyclic) peptide compounds.
- the number of unnatural amino acids, N-substituted amino acids, or amino acids having an aromatic ring contained in the peptide compound contained in the library or the peptide compound encoded by the nucleic acid contained in the library is used.
- the average can be adjusted by the appearance frequency of the codon assigned to the amino acid when the DNA library is synthesized. For example, in a library of peptides containing variable positions of 10 amino acid residues, assuming that the amino acids are selected from 20 kinds, by setting 10 kinds of N-substituted amino acids in the 20 kinds, the codon unit of the N-substituted amino acids can be obtained. Can be synthesized so that each variable point occupies 10/20 (50%). In this way, the average of the ratio of the number of unnatural amino acids, the number of N-substituted amino acids, the number of aromatic rings, or the number of amino acids having an aromatic ring in the present specification may be calculated.
- the phrase “consisting essentially of” means the theory of variations of the peptide compounds contained in the library and/or the nucleic acids encoding them.
- the ratio of the theoretical number of variations of the listed peptide compounds and/or nucleic acids encoding them to the above total number is 90% or more, 93% or more, 95% or more, 97% or more, 98% Or more, or 99% or more, but not limited to these.
- a codon table in which one set of codon units corresponds to one type of amino acid is used, and a peptide having 11 residues is selected from a base sequence in which the codon units are randomly arranged.
- the theoretical number of variations is 20 10 .
- the library herein comprises peptide compounds, and/or nucleic acids encoding them, as well as other components necessary for screening peptide compounds capable of specifically binding to a target molecule. You can stay. Further, other components may be contained within a range not negatively affecting the effect of the library in the present specification.
- the library in the present specification includes a plurality of peptide compounds or nucleic acids encoding them.
- the number of variations (types) of peptide compounds contained in the library is called “diversity”.
- the diversity of the peptide compounds contained in the library in the present specification, or the nucleic acids encoding these is not particularly limited, but 10 3 or more, 10 4 or more, 10 5 or more, 10 6 or more, 10 7 or more, 10 8 or more, Examples include 10 9 or more, 10 10 or more, 10 11 or more, and 10 12 or more. These varieties are not limited to actually measured values and may be theoretical values.
- the average molecular weight of the peptide portion other than the nucleic acid junction of the peptide compound contained in the library or encoded by the nucleic acid contained in the library, or the molecular weight of the cyclic portion of the cyclic peptide compound is used. May average from 500 to 2000.
- a display library is a library in which a phenotypic peptide is associated with an RNA or a DNA encoding the genotypic peptide.
- This library can be used to identify peptide compounds that can specifically bind to a target molecule. For example, by contacting the library with a desired fixed target and washing away molecules that do not bind to the target, it is possible to concentrate the peptide that binds to the target (panning method). The sequence of the peptide bound to the target can be clarified by analyzing the genetic information associated with the selected peptide through such a process.
- a method utilizing the fact that the antibiotic puromycin, which is an analog of aminoacyl-tRNA, is nonspecifically linked to a protein during mRNA translation elongation by the ribosome is mRNA display (Proc Natl Acad Sci USA. 1997;94:12297- 302.RNA-peptide fusions for the in vitro selection of peptides and proteins.Roberts RW, Szostak JW.) or in vitro virus (FEBS Lett. 1997;414:405-8.
- In vitro virus bonding of mRNA bearing puromycin at the 3'-terminal end to the C-terminal end of its encoded protein on the ribosome in vitro. Nemoto N, Miyamoto-Sato E, Husimi Y, Yanagawa H.).
- a spacer such as puromycin is bound to the 3'end of a library of mRNA obtained by transcription from a DNA library containing a promoter such as T7 promoter, and the mRNA is translated into a protein in a cell-free translation system, puromycin becomes a ribosome. Is mistakenly taken as an amino acid into a protein and linked to the mRNA and the protein encoded by the mRNA, resulting in a library in which the mRNA and its product are associated with each other. Since this process does not involve transformation of E. coli and the like, high efficiency is realized and a large-scale display library can be constructed.
- the sequence of the bound protein can be clarified by synthesizing cDNA from mRNA, which is a tag containing gene information of the molecule selected and enriched by panning, PCR-amplified, and analyzing the nucleotide sequence.
- a display library utilizing a cell-free translation system is a cDNA display library (Nucleic Acids Res. 2009; 37, which is a library consisting of a cDNA encoding a peptide in which a peptide-puromycin complex is bound).
- CISdisplay Invitro selection of peptides from libraries of protein-DNA complexes. Odegrip R, Coomber D, Eldridge B, HedererR, Kuhlman PA, Ullman C, FitzGerald K, McGregor.D. In addition, in vitro compartmentalization (Nat Biotechnol. 1998;16:652-6. Man- made cell-like compartmentsfor molecular evolution.Tawfik DS, Griffiths AD.) are also known. The above method can be appropriately used by using a known method.
- Nucleic acid library The "nucleic acid” in the present invention can also include deoxyribonucleic acid (DNA), ribonucleic acid (RNA), or a nucleotide derivative having an artificial base. It may also include peptide nucleic acid (PNA).
- the nucleic acid of the present invention may be any of these nucleic acids or a hybrid thereof as long as the desired genetic information is retained. That is, a chimeric nucleic acid in which different nucleic acids such as a DNA-RNA hybrid nucleotide and DNA and RNA are linked in a single strand is also included in the nucleic acid of the present invention.
- a nucleic acid library can be obtained by synthesizing bases at positions where amino acid residues are not fixed on the peptide sequence. For example, for a DNA library, A, T, G, C, and for an RNA library, each of A, U, G, and C is a multiple of 3 repeated 4 nucleotides (N).
- the third letter is synthesized as a mixture of two bases such as N, the third letter is W, M, K, S, etc., and if the number of introduced amino acids is 16 or less, the third letter should be one type of base. There is also a way to do it.
- the appearance frequency of amino acid residues can be freely adjusted by preparing codon units corresponding to the three letters of the codon, mixing them at an arbitrary ratio and using them for synthesis.
- nucleic acid libraries can be translated using a cell-free translation system.
- a sequence encoding a spacer downstream of the target nucleic acid examples include, but are not limited to, sequences including glycine and serine.
- a linker formed of RNA, DNA, a polymer of hexaethylene glycol (spc18) (for example, 5 polymers) is included between the nucleic acid library and a compound that is incorporated into a peptide during translation by the ribosome, such as puromycin or a derivative thereof.
- spc18 hexaethylene glycol
- the library according to the present disclosure can be produced according to the above-described method for producing a peptide compound according to the present disclosure, and can be appropriately combined with a known method.
- the cell-free translation system of the present disclosure described above can be used to produce a library of peptide compounds of the present disclosure. That is, the method for producing a library according to the present disclosure may include a step of synthesizing a peptide compound using the cell-free translation system according to the present disclosure.
- the exemplification, the preferred range, and the aspect described in the cell-free translation system of the present disclosure can be directly applied to the method for producing a library of the present disclosure.
- the mRNA display library can be produced as follows in a non-limiting aspect. First, a library of DNA in which a desired sequence is arranged downstream of a promoter such as T7 promoter is chemically synthesized, and this is used as a template to form a double-stranded DNA by a primer extension reaction. Using this as a template, RNA polymerase such as T7 RNA polymerase is used to transcribe it into mRNA. A linker (spacer) in which an antibiotic puromycin or the like which is an analog of aminoacyl-tRNA is connected to the 3'end of this RNA is bound.
- a promoter such as T7 promoter
- the library in the present disclosure is a peptide compound containing a peptide compound containing at least one, two or more, or three or more amino acids selected from the group consisting of the amino acids described herein. It may be a library.
- a method for producing such a library may include a step of translationally synthesizing a peptide compound in a cell-free translation system including the following (i) and (ii): (i) tRNA having at least one, two or more, or three or more amino acids selected from the group consisting of the amino acids described in the present specification; (ii) a nucleic acid library encoding the peptide compound library,
- the nucleic acid library may include a nucleic acid containing at least one codon corresponding to the anticodon of the tRNA.
- the reaction solvent used for peptide synthesis and solid phase synthesis was used for peptide synthesis (purchased from Watanabe Kagaku and Wako Pure Chemical Industries). For example, DCM, DMF, NMP, 2% DBU in DMF, 20% piperidine in DMF and the like. Further, in the reaction in which water was not added as a solvent, a dehydrated solvent, a super dehydrated solvent, or an anhydrous solvent (purchased from Kanto Kagaku, Wako Pure Chemical Industries, Ltd.) was used.
- the LC/MS analysis conditions are as follows.
- Example 1 Chemical Synthesis of Peptide Compound According to the peptide synthesis method by the Fmoc method described in WO2013/100132, the peptide was extended according to the following basic route. That is, 1) the carboxylic acid of the Asp side chain was carried on 2-chlorotrityl resin, but the peptide extension reaction by the Fmoc method from the N terminus of Asp was performed, 2) the process of excising the peptide from 2-chlorotrityl resin, 3 ) Amide cyclization by condensation of the carboxylic acid of the Asp side chain generated outside the 2-chlorotrityl resin by the excision process and the amino group at the N-terminal (triangle unit) of the peptide chain, 4) side chain function contained in the peptide chain It is a 5-step process including deprotection of the protecting group of the group, 5) purification of the compound by prepparative HPLC.
- Fmoc-nPrGly-OH, Fmoc-MePhe(3-Cl)-OH, Fmoc-MeAla(4-Thz)-OH and the like were synthesized by the method described in WO2013/100132.
- Fmoc-Thr(THP)-OH (Compound aa01), Fmoc-nBuGly-OH (Compound aa02), Fmoc-Ser(iPen)-OH (Compound aa04), Fmoc-MeHph-OH (Compound aa05), Fmoc- Ser(3-F-5-Me-Pyr)-OH (Compound aa12), Fmoc-Ser(Ph-3-Cl)-OH (Compound aa16), Fmoc-Nle(6-OTHP)-OH (Compound aa17) , Fmoc-Ser(EtOTHP)-OH (Compound aa22), Fmoc-MeSer(EtOTHP)-OH (Compound aa27), Fmoc-Ser(S-2-PrOTHP)-OH (Compound aa34), Fmoc-MeSer(S-).
- BuOTHP -OH (Compound aa64), Fmoc-Hnl(7-F3-6-OH)-OH (Compound aa68), Fmoc-Ser(1-CF3-EtOH)-OH (Compound aa70), Fmoc-MeSer(1).
- a sequence containing an amino acid having a tertiary alcohol in the side chain for example, Ser(tBuOH), MeSer(tBuOH), Ser(2-Me-2-BuOH), MeSer(2-Me-2-BuOH), etc.
- the peptide was extended without any problem even if an Fmoc amino acid in which the tertiary alcohol portion of the side chain was not protected with THP was used (eg, Fmoc-Ser(tBuOH)-OH (compound aa53), Fmoc-Ser(2-Me-2). -BuOH)-OH (compound aa61) was used).
- An amino acid having a structure in which the side chain is a secondary alcohol and the carbon to which the alcohol is bonded has a CF3 group bonded thereto eg, Hnl(7-F3-6-OH), Ser(1-CF3-EtOH) ) Etc.
- the peptide was extended (eg, Fmoc-Hnl(7-F3-6-OH)-OH) without any problem even if an Fmoc amino acid that does not protect the side chain secondary alcohol site with THP was used.
- Compound aa68 Fmoc-Ser(1-CF3-EtOH)-OH (Compound aa70) and the like were used).
- a saturated aqueous sodium chloride solution (30 mL) was added to wash the organic layer, and the aqueous layer was extracted with ethyl acetate (30 mL). All the obtained organic layers were mixed, and this was further washed twice with a saturated aqueous sodium chloride solution (30 mL). The organic layer was dried over sodium sulfate, and the solvent was evaporated under reduced pressure to give a crude product (9.3 g). Of the obtained crude product, 4.65 g was dissolved in tetrahydrofuran (THF) (30 mL), and then 1.0 M phosphate buffer (30 mL) adjusted to pH 8.0 was added. The mixture was stirred at 50 degrees for 4 hours.
- THF tetrahydrofuran
- N-(tert-butoxycarbonyl)-O-isopentyl-L-serine (Boc-Ser(iPen)-OH) (183 g). ..
- the obtained N-(tert-butoxycarbonyl)-O-isopentyl-L-serine (Boc-Ser(iPen)-OH) (175 g) was dissolved in 1,4-dioxane (500 mL), and concentrated hydrochloric acid ( (300 mL) was added, and the mixture was stirred at room temperature for 16 hours.
- the reaction mixture was concentrated under reduced pressure, the obtained crude product (122 g) was dissolved in 1,4-dioxane/water (1000 mL/1000 mL), potassium carbonate (239 g, 1.73 mol) and N- (9-Fluorenylmethoxycarbonyloxy)succinimide (Fmoc-OSu) (234 g, 694.36 mmol) was added, and the mixture was stirred at room temperature for 16 hours. Then, the reaction solution was washed with hexane three times. After washing, a hydrochloric acid aqueous solution (6M) was added to adjust the pH of the aqueous layer to 2 to 3, and the mixture was extracted twice with ethyl acetate.
- 1,4-dioxane/water 1000 mL/1000 mL
- potassium carbonate 239 g, 1.73 mol
- N- (9-Fluorenylmethoxycarbonyloxy)succinimide Fm
- Triethylsilane (Et 3 SiH) and trifluoroacetic acid (TFA) (700 mL) were added, and the mixture was stirred at 25° C. for 16 hr.
- the reaction mixture was concentrated under reduced pressure, aqueous potassium carbonate solution was added to the obtained residue, and the mixture was washed with petroleum ether.
- reaction solution was cooled to ⁇ 60° C. or lower, and a THF solution of LiAlH 4 (lithium aluminum hydride) (2.5 M, 40 mL, 100 mmol) was added to the reaction solution ⁇ 20.
- LiAlH 4 lithium aluminum hydride
- the solution was added dropwise over 5 minutes so that the temperature did not exceed °C.
- the reaction solution was stirred at ⁇ 78° C. for 3 hours, and ethyl acetate (69 mL) was added so that the reaction solution did not exceed ⁇ 20° C.
- the mixture was further stirred for 1 hour using an ice bath, water (18 mL) was added, and the mixture was stirred for 15 minutes at room temperature.
- reaction solution was stirred at room temperature for another hour, and water (125 mL) was added.
- the obtained mixed liquid was washed with t-butyl methyl ether (TBME), and the organic layer was extracted with water.
- TBME t-butyl methyl ether
- the obtained organic layers were combined, washed three times with a solution obtained by diluting saturated saline twice, and then washed twice with saturated saline.
- N-succinimidyl carbonate 9-fluorenylmethyl carbonate (Fmoc-OSu) (7.86 g, 23.3 mmol) was added to the reaction solution at room temperature, and the mixture was stirred at room temperature.
- water (56.2 mL) was added to the reaction solution at room temperature, and the mixture was washed twice with a 25% t-butyl methyl ether (MTBE)/hexane solution.
- MTBE 25% t-butyl methyl ether
- the obtained residue was dissolved in tetrahydrofuran (15 mL), dihydropyran (5.41 mL, 56.8 mmol) was added, and the mixture was stirred at 50°C for 1 hr. Ethyl acetate was added to the reaction solution at room temperature, and the organic layer was washed twice with saturated saline, then, the obtained organic layer was dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure. The obtained residue was dissolved in tetrahydrofuran (30 mL), phosphate buffer (30 mL) with pH adjusted to 8 was added, and the mixture was stirred at 50° C. for 3 hours.
- the reaction solution was extracted twice with ethyl acetate at room temperature, the obtained organic layer was washed with saturated brine, the organic layer was dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure.
- the obtained residue was dissolved in diethyl ether (50 mL), heptane (50 mL) was added thereto to precipitate a crude product, which was concentrated under reduced pressure to remove diethyl ether, and the remaining supernatant solution was added. It was removed by decantation. A crude product was obtained by repeating this operation twice.
- the obtained crude product was dissolved in tert-butyl methyl ether (150 mL), 0.05M phosphoric acid aqueous solution (150 mL) was added, and the mixture was stirred at room temperature for 1 hr. The organic layer was collected, and the aqueous layer was extracted with ethyl acetate.
- Triethylsilane (24.4 g, 210 mmol) and trifluoroacetic acid (450 mL) were added to the solution (450 mL), and the mixture was stirred at room temperature for 48 hours.
- the reaction solution was evaporated under reduced pressure, dried under reduced pressure using a vacuum pump, redissolved in an aqueous potassium carbonate solution, washed with diethyl ether, and concentrated hydrochloric acid was added until pH 2 was reached. Extracted.
- reaction solution was evaporated under reduced pressure, further dried under reduced pressure with a vacuum pump, redissolved in an aqueous potassium carbonate solution, washed with diethyl ether, and concentrated hydrochloric acid was added until pH 2 was reached. Extracted. The obtained organic layers were mixed and dried over anhydrous sodium sulfate, the solvent was evaporated under reduced pressure, and the residue was dried under reduced pressure with a vacuum pump to give N-((benzyloxy)carbonyl)-O-((S)-2.
- Methyl N-((benzyloxy)carbonyl)-O-((R)-2-(benzyloxy)propyl)-L-serinate (Compound aa42, Cbz-Ser(R-2-PrOBn)-OMe)( A solution of 25.0 g, 62.3 mmol) in 2-propanol/tetrahydrofuran (1.50 L/150 mL) was added at room temperature, and the mixture was stirred for 5 hours. A 2M aqueous hydrochloric acid solution was added until the pH reached 2, the organic layer was removed, and the aqueous layer was extracted 3 times with ethyl acetate.
- the pH of the aqueous layer was adjusted to 1 by adding a 2 M aqueous hydrochloric acid solution (26.9 mL, 53.8 mmol) to the aqueous layer, and then extracted with t-butyl methyl ether (60 mL) three times.
- the organic layer is dried over anhydrous magnesium sulfate, filtered, and t-butyl methyl ether is concentrated under reduced pressure.
- the residue obtained is purified by reverse phase column chromatography (0.1% formic acid aqueous solution/0.1 formic acid acetonitrile solution).
- the obtained residue was dissolved in tetrahydrofuran (24.0 mL), 1.0 M phosphate buffer (24.0 mL) adjusted to pH 6.8 was added, and the mixture was stirred at 50° C. for 3 hr.
- the reaction solution was diluted with ethyl acetate (45 mL), washed with saturated brine (45 mL) three times, and the organic layer was dried over anhydrous sodium sulfate.
- a toluene/methanol (90 mL/60 mL) solution was added to the residue obtained by concentrating the reaction solution under reduced pressure using a rotary evaporator, and after dissolving it, a 2M (trimethylsilyl)diazomethane/hexane solution (24.26) was added. (mL, 48.5 mmol) was added dropwise at room temperature, and the mixture was stirred for 6 hours.
- the reaction solution was concentrated under reduced pressure using a rotary evaporator to give a crude product of methyl (S)-2-((tert-butoxycarbonyl)amino)-6-hydroxyhexanoate (Boc-Nle(6-OH)- OMe) (13 g) was obtained.
- reaction solution was concentrated under reduced pressure with a rotary evaporator to remove tetrahydrofuran, ethyl acetate and a 1N aqueous hydrochloric acid solution were added, and the mixture was extracted twice with ethyl acetate. The organic layer was washed with saturated brine, dried over anhydrous magnesium sulfate, filtered, and concentrated under reduced pressure.
- the obtained residue was purified by reverse phase column chromatography (0.1% formic acid aqueous solution/0.1% formic acid acetonitrile solution) and normal phase column chromatography (dichloromethane/methanol), and then further reverse phase column chromatography ( Fractions obtained by purification with 10 mM ammonium acetate aqueous solution/methanol) were collected, methanol was evaporated under reduced pressure, extraction was performed twice with ethyl acetate, and the extract was washed with saturated potassium hydrogen sulfate solution and saturated saline.
- the obtained residue was dissolved in dichloromethane (60 mL) and heptane (60 mL), and dichloromethane was concentrated under reduced pressure to precipitate an oily crude product.
- the heptane solution was removed by decantation. This operation was repeated twice.
- the obtained crude product was dissolved in ethyl acetate and washed twice with a 0.05 M phosphoric acid aqueous solution and once with a saturated saline solution.
- N,N-diisopropylethylamine (DIPEA) 127 mL
- DIPEA 9H-fluoren-9-yl)methyl carbonochloride
- Fmoc-Cl 9H-fluoren-9-ylmethyl carbonochloride
- 1,4-dioxane solution 145 mL was added dropwise, and the reaction mixture was stirred at 0° C. for 1 hr 30 min.
- the solvent of the reaction solution was evaporated under reduced pressure, diluted with ethyl acetate, and then washed twice with water and a saturated aqueous solution of ammonium chloride and twice with a saturated saline solution.
- N-ethyl-isopropylpropan-2-amine DIPEA
- DIPEA N-ethyl-isopropylpropan-2-amine
- DIPEA 1,4-dioxane
- DIPEA N-ethyl-isopropylpropan-2-amine
- Succinimide Fmoc-OSu (22.3 g, 231 mmol) was added, and the mixture was stirred at 25° C. for 16 hours.
- boron trifluoride diethyl ether complex (BF 3 ⁇ OEt 2 ) (0.580 mL, 4.57 mmol) was added, and the mixture was stirred at room temperature for 24 hours.
- a saturated aqueous ammonium chloride solution was added to the reaction solution, the mixture was stirred at room temperature for 40 minutes, and then extracted twice with dichloromethane (DCM).
- DCM dichloromethane
- the reaction solution was added to a mixed solution of ethyl acetate (300 mL), water (60 mL) and 1N aqueous hydrochloric acid solution (4.8 mL), and the obtained organic layer was evaporated under reduced pressure to obtain a crude product.
- the above two crude products were combined and purified by reverse phase column chromatography (0.1% formic acid aqueous solution/0.1% formic acid acetonitrile solution) to give O-(2-(methylamino)-2-oxoethyl).
- -N-Trityl-L-serine (Trt-Ser(NMe-Aca)-OH) (2.5 g) was obtained.
- N-Ethyl-N-isopropylpropan-2-amine (DIPEA) (2.231 mL, 12.77 mmol) in 1,4-dioxane (14.0 mL) was added at room temperature, and then N- (9-Fluorenylmethoxycarbonyloxy)succinimide (Fmoc-OSu) (1.723 g, 5.11 mmol) was added at room temperature, and the mixture was stirred for 30 minutes.
- the reaction solution was added to a mixed solution of ethyl acetate (150 mL), water (30 mL) and 1N hydrochloric acid aqueous solution (3.0 mL), and the obtained organic layer was evaporated under reduced pressure.
- a tetrahydrofuran solution (134 mL) of 762 mmol) was added at room temperature and stirred for 5 hours.
- a 6N aqueous hydrochloric acid solution was added to the reaction solution until the pH reached 3, the organic solvent was removed under reduced pressure, and the aqueous layer was extracted with ethyl acetate.
- N-(9-Fluorenylmethoxycarbonyloxy)succinimide (Fmoc-OSu) (388 mg, 1.15 mmol) was added again to the reaction solution at 0° C., and the mixture was stirred for 40 minutes.
- Water and t-butyl methyl ether (TBME) were added to the reaction solution, the obtained aqueous layer was acidified with a 1N aqueous hydrochloric acid solution, and extracted three times with ethyl acetate. The obtained organic layers were combined, washed with saturated brine, dried over anhydrous magnesium sulfate and filtered, and then the solvent was evaporated under reduced pressure.
- TMS diazomethane 2-methylsilyldiazomethane (2.62 mL, 5.23 mmol) was again added dropwise to the reaction solution at 0 degrees, and then 2.0M trimethylsilyldiazomethane (TMS diazomethane) was added in hexane.
- the solution (3.93 mL, 7.85 mmol) was further added dropwise at 0°C.
- the reaction was stopped by adding acetic acid (2.0 mL), 50% saturated aqueous sodium hydrogen carbonate solution and ethyl acetate were added, and the mixture was washed with ethyl acetate.
- a diethyl ether solution of diazomethane separately prepared was added dropwise to the reaction solution while cooling using an ice bath prepared with ice and sodium chloride, and the mixture was stirred for 30 minutes.
- Acetic acid (0.88 mL, 15.37 mmol) was added to the reaction mixture, the mixture was stirred for 10 minutes, ethyl acetate and water were added, the mixture was extracted with ethyl acetate, and the obtained organic layer was washed with saturated brine, It was dried over anhydrous sodium sulfate and filtered.
- Diazomethane Diethyl Ether Solution 1-Methyl-3-nitro-1-nitrosoguanidine (5.0 g, purchased from Tokyo Kasei or Fuji Film Wako Pure Chemical Industries) containing about 50% water was added with diethyl ether (17. 0 mL) was added, and a 50% aqueous potassium hydroxide solution (9.5 mL) was added dropwise at 0°C. After stirring the reaction solution for 1 hour, the yellow diethyl ether layer was separated using a Pasteur with a tanned tip, and the obtained solution was used as it was in the above reaction.
- the obtained organic layer was washed with saturated saline, dried over anhydrous sodium sulfate, and filtered.
- Water 50 mL was added to the reaction solution, and the mixture was extracted with t-butyl methyl ether/hexane (40 mL/10 mL).
- the obtained organic layer was washed with saturated saline, dried over anhydrous sodium sulfate, and filtered.
- methyltriphenylphosphonium bromide 562 mg, 1.57 mmol
- diethyl ether solution 3.57 mmol
- potassium 2-methylpropane-2-olate tBuOK
- 0.57 mmol was added and the mixture was stirred at room temperature for 2 hours.
- a separately prepared aldehyde intermediate (185 mg, 0.786 mmol) in diethyl ether (0.79 mL) was added to the reaction solution, and the mixture was stirred at room temperature for 1 hour.
- a 10% aqueous ammonium chloride solution was added to the reaction solution, followed by extraction with ethyl acetate, and the obtained organic layer was washed with water.
- Second generation Hoveyda-Grubbs catalyst (0.94 g, 1.50 mmol) was added to the solution and stirred at 60° C. overnight. After cooling to room temperature, it was filtered through Celite, and the filtrate was concentrated under reduced pressure. The obtained residue was purified by reverse phase silica gel column chromatography (0.1% formic acid aqueous solution/0.1% formic acid acetonitrile solution) to obtain a crude product (7.58 g).
- the obtained crude product (7.58 g) was dissolved in methanol (186 mL), 10% palladium carbon (1.6 g) was added, and the mixture was stirred under a hydrogen atmosphere at room temperature for 2 days.
- the reaction solution was filtered through Celite, and the filtrate was concentrated under reduced pressure.
- the obtained residue was dissolved in methanol (34.9 mL), saturated aqueous sodium hydrogen carbonate solution (34.9 mL), N-(9-fluorenylmethoxycarbonyloxy)succinimide (Fmoc-OSu) (7.07). g, 20.95 mmol) was added and the mixture was stirred at room temperature for 2 hours.
- the insoluble material was filtered off, the filtrate was acidified with a 1M aqueous hydrochloric acid solution, and then extracted with ethyl acetate. The organic layer was washed with saturated brine, dried over anhydrous magnesium sulfate, filtered, and concentrated under reduced pressure.
- Tetrahydrofuran (THF) of (S)-4,4-difluoropyrrolidine-1,2-dicarboxylic acid 2-methyl 1-allyl (compound aa123) (6.81 g, 27.3 mmol) obtained as described above
- a lithium borohydride/tetrahydrofuran (THF) solution (10.24 mL, 41.0 mmol) was added dropwise to the solution (82.0 mL) at 0° C., and the mixture was stirred at room temperature for 2 hours.
- lithium hydroxide monohydrate (1.89 g, 44.9 mmol) was added at room temperature to an aqueous solution (46.8 mL) of calcium chloride (18.7 g, 168 mmol), and at room temperature. Stir for 5 minutes.
- (2S,4R)-1-((benzyloxy)carbonyl)-4-hydroxypyrrolidine-2-carboxylic acid (Cbz-Hyp-OH) (12.0 g, 45 at ⁇ 10° C.
- Step 1 Methyl 3-((2-(tert-butoxy)-2-oxoethyl)amino)propanoate (31.0) prepared by the method described in the literature (Journal of Medicinal Chemistry (1991), 34(4), 1297-1301). g, 142.683 mmol) and N-(9-fluorenylmethoxycarbonyloxy)succinimide (Fmoc-OSu) (48.13 g, 142.683 mmol) in 1,4-dioxane solution (200 mL) was carbonated. An aqueous solution (200 mL) of sodium (30.25 g, 285.366 mmol) was added at room temperature, and the mixture was stirred for 16 hours.
- Step 2 After adding water (250 mL) and isopropanol (1000 mL) to calcium chloride (84.54 g, 761.761 mmol) and lithium hydroxide monohydrate (8.52 g, 203.136 mmol), The above crude product methyl 3-((((9H-fluoren-9-yl)methoxy)carbonyl)(2-(tert-butoxy)-2-oxoethyl)amino)propanoate (62 g, 50.784) A tetrahydrofuran solution (250 mL) of (mmol) was added at 0° C., and the mixture was stirred for 16 hours.
- Step 3 Under a nitrogen atmosphere, the above crude product 3-((((9H-fluoren-9-yl)methoxy)carbonyl)(2-(tert-butoxy)-2-oxoethyl)amino)propanoic acid (14 g, 32.904 mmol), N-hydroxyphthalimide (5.90 g, 36.194 mmol), N,N'-dicyclohexylcarbodiimide (DCC) (7.47 g, 36.194 mmol) in tetrahydrofuran (140 mL) was stirred at room temperature for 16 hours.
- N-hydroxyphthalimide 5.90 g, 36.194 mmol
- DCC N,N'-dicyclohexylcarbodiimide
- Step 4 Under a nitrogen atmosphere, trihydrate nickel bromide (II) (NiBr 2 ⁇ 3H 2 O) (5.73 g, 21.031mmol) and 4,4'-di -tert- butyl-2,2'- A solution of bipyridyl (dtbbpy) (5.64 g, 21.031 mmol) in dimethylacetamide (DMA) (200 mL) was stirred at 50° C. for 3 hours and then cooled to room temperature to obtain a nickel catalyst solution. Under a nitrogen atmosphere, zinc powder (22.93 g, 350.509 mmol), crude product 3-((((9H-fluoren-9-yl)methoxy)carbonyl)(2) obtained as described above.
- II trihydrate nickel bromide
- DMA dimethylacetamide
- Step 5 The above tert-butyl N-(((9H-fluoren-9-yl)methoxy)carbonyl)-N-(3-methoxy-4-(methylcarbamoyl)phenethyl)glycinate (14.0 g, 25.705 mmol) 4N hydrochloric acid/1,4-dioxane solution (65 mL) was stirred at room temperature for 48 hours. The reaction mixture was concentrated under reduced pressure, and the obtained residue was purified by reverse phase column chromatography (water/acetonitrile) to give N-(((9H-fluoren-9-yl)methoxy)carbonyl)-N.
- Step 1 Commercially available (S)-4-((((9H-fluoren-9-yl)methoxy)carbonyl)amino)-5-(tert-butoxy)-5-oxopentanoic acid (Fmoc-Glu-OtBu) under nitrogen atmosphere.
- Step 2 Under a nitrogen atmosphere, nickel (II) bromide trihydrate (NiBr 2 .3H 2 O) (141 mg, 0.526 mmol) and 4,4′-di-tert-butyl-2,2′-bipyridyl were used.
- a dimethylacetamide (DMA) solution (5 mL) of (dtbbpy) (140 mg, 0.526 mmol) was stirred at 50° C. for 3 hours and then cooled to room temperature to obtain a nickel catalyst solution.
- DMA dimethylacetamide
- Zinc powder (617 mg, 8.75 mmol) under a nitrogen atmosphere, (((9H-fluoren-9-yl)methoxy)carbonyl)-L-glutamic acid 5-(1,3-di) obtained as described above.
- DMA dimethylacetamide
- Step 3 (S)-2-((((9H-fluoren-9-yl)methoxy)carbonyl)amino)-4-(3-methoxy-4-(methylcarbamoyl)phenyl)butanoic acid tert obtained as described above -Chlorotrimethylsilane (TMS-Cl) (1.80 g, 16.568 mmol) was added to a trifluoroethanol (TFE) solution (30 mL) of butyl (3.00 g, 5.508 mmol) at room temperature. Stir for 3 hours.
- TFE trifluoroethanol
- Aqueous ammonium chloride solution was added to the reaction solution, the mixture was extracted twice with ethyl acetate, and the obtained organic layer was concentrated under reduced pressure.
- An aqueous solution of sodium hydrogen carbonate was added to the residue to dissolve it, and the mixture was washed 3 times with hexane/ethyl acetate (5/1).
- the organic layer was washed with water and saturated brine, dried over anhydrous sodium sulfate and filtered, and the filtrate was concentrated under reduced pressure.
- (2S)-2-((((9H-fluoren-9-yl)methoxy)carbonyl)amino)-3-(4-(methyl((tetrahydro-2H-pyran-2-yl)oxy)carbamoyl)phenyl) Propanic acid compound aa139, Fmoc-Phe(4-CONMeOTHP)-OH
- (2S)-2-((((9H-fluoren-9-yl)methoxy)carbonyl)amino)-3-(3-methoxy- 4-(((tetrahydro-2H-pyran-2-yl)oxy)carbamoyl)phenyl)propanoic acid compound aa140, Fmoc-Phe(3-OMe-4-CONHOTHP)-OH was synthesized according to the following scheme.
- Zinc powder (17.13 g, 261.856 mmol) under nitrogen atmosphere, (((9H-fluoren-9-yl)methoxy)carbonyl)-L-aspartic acid 1-methyl 4-(synthesized as above. 1,3-dioxoisoindoline-2-yl) (26.94 g, 52.371 mmol), tert-butyl 4-iodo-2-methoxybenzoate (35 as the crude product synthesized as described above).
- Step 1 Under a nitrogen atmosphere, an aqueous solution (100 mL) of commercially available (S)-morpholine-3-carboxylic acid (CAS. No. 741288-31-3) (14.0 g, 107 mmol) was added at room temperature to sodium hydrogen carbonate ( 20.26 g, 241.172 mmol) was added. A 1,4-dioxane solution (100 mL) of allyl chloroformate (Alloc-Cl) (17.44 g, 120.53 mmol) was added dropwise to the reaction solution at room temperature, and the mixture was stirred for 2 hours. Water was added to the reaction solution, followed by extraction with ethyl acetate.
- Step 2 A solution of 3-methyl-4-allyl (S)-morpholine-3,4-dicarboxylate (19.5 g, 85.067 mmol) obtained as described above in a tetrahydrofuran (THF) solution (283 mL) was added at 0 degree. Lithium borohydride (5.56 g, 255.233)/tetrahydrofuran (THF) solution was added dropwise, and the mixture was stirred at room temperature for 2 hours. Water and a 2N hydrochloric acid aqueous solution were added to the reaction solution at 0° C., followed by extraction with ethyl acetate.
- THF tetrahydrofuran
- Step 3 In a nitrogen atmosphere, (S) was synthesized by the above-mentioned allyl (R)-3-(hydroxymethyl)morpholine-4-carboxylate (7.0 g, 34.8 mmol) and the method described in the patent (WO2015/179823).
- -Aziridin-1,2-dicarboxylic acid 2-methyl-1-((9H-fluoren-9-yl)methyl) (7.50 g, 23.2 mmol) in dichloromethane solution (47 mL) at 0°C.
- a dichloromethane solution of boron fluoride-ethyl ether complex (1.48 g, 10.45 mmol) was added dropwise, and the mixture was stirred for 2 hours.
- Step 4 Under a nitrogen atmosphere, lithium hydroxide monohydrate (0.91 g, 37.999 mmol) was added at room temperature to an aqueous solution (50 mL) of calcium chloride (15.87 g, 142.974 mmol), and at room temperature. Stir for 10 minutes.
- Resin synthesis used for peptide synthesis by peptide synthesizer was synthesized as follows. 2-Chlorotrityl chloride resin (100-200 mesh, 1% DVB) was purchased from Watanabe Chemical Industry and Chem-Impex.
- the polymer or resin portion when a polymer or resin and a compound are bound to each other, the polymer or resin portion may be indicated by a circle.
- the reaction site of the resin site it may be connected to a circle to indicate the chemical structure of the reaction site.
- the 2-chlorotrityl group on the resin is bonded to the side chain carboxylic acid of Asp via an ester bond. It shows how they are doing.
- a 5% aqueous sodium hydrogen carbonate solution was added to the reaction solution, and the mixture was washed twice with t-butyl methyl ether (TBME).
- TBME t-butyl methyl ether
- the pH of the obtained aqueous layer was acidified with a 6 M aqueous hydrochloric acid solution and then extracted with ethyl acetate.
- the obtained organic layer was washed twice with 50% saline, dried over anhydrous magnesium sulfate, and the obtained mixture was filtered and concentrated under reduced pressure.
- Peptide solid phase synthesis by an automatic synthesizer was performed by Fmoc method using a peptide synthesizer (Multipep RS; manufactured by Intavis) by the method described in WO2013/100132. The detailed procedure of the operation was according to the manual attached to the synthesizer.
- 2-Chlorotritylresin eg (S)-3-((((9H-fluoren-9-yl)methoxy)carbonyl)carbonyl having a side chain carboxylic acid moiety of aspartic acid with N-terminal Fmoc protected attached to the synthesizer )Amino)-4-oxo-4-(piperidin-1-yl)butanoic acid-2-chlorotrityl resin (compound pd01, Fmoc-Asp(O-Trt(2-Cl)-resin)-pip))(1 100 mg per column), various Fmoc-amino acids (0.6 mol/L) and 1-hydroxy-7-azabenzotriazole (HOAt) or oxyma (0.375 mol/L) N-methyl-2-pyrrolidone A (NMP) solution and a N,N-dimethylformamide (DMF) solution of diisopropylcarbodiimide (DIC)
- Fmoc-Thr(THP)-OH (compound aa01) is made to coexist with oxyma in an NMP solution, and molecular sieves 4A1/8 (Wako Pure Chemical Industries) or molecular sieves 4A1/16 (Wako Pure Chemical Industries) is added to set. did.
- molecular sieves 4A1/8 (Wako Pure Chemical Industries) or molecular sieves 4A1/16 (Wako Pure Chemical Industries) is added to set.
- an amino acid having a THP group as a protecting group of the amino acid side chain eg, Fmoc-Ser(EtOTHP)-OH (Compound aa22) is also oxyma in NMP solution.
- a peptide containing an amino acid having an Alloc group in its side chain is Zinc chloride (1.0 equivalent to the number of moles on the resin used), tetrakis(triphenylphosphine)palladium(0)(Pd(PPh 3 ) 4 ) (based on the number of moles on the resin used) 1.0 equivalent), poly(methylhydrosiloxane) (PMHS, CAS#63148-57-2) (0.9 M relative to the number of moles on the resin used), tetrahydrofuran (number of moles on the resin used) 0.1 M) was used to deprotect the Alloc group by stirring at room temperature for 3 hours and 30 minutes or more, and then 0.05 M tetramethylammonium hydrogensulfate/1,1M prepared in the reaction solution was prepared.
- Zinc chloride 1.0 equivalent to the number of moles on the resin used
- tetrakis(triphenylphosphine)palladium(0)(Pd(PPh 3 ) 4 )
- HFIP 1,3,3,3-hexafluoroisopropyl alcohol
- TIPS triisopropylsilane
- DIPEA diisopropylethylamine
- TMDS tetramethyldisiloxane
- Example 2 Evaluation of membrane permeability of an amino acid having a donor in the side chain and an intramolecular hydrogen bond in the donor moiety 2-1
- Membrane permeability comparison of amino acids having a donor in the side chain and those not having an intramolecular hydrogen bond in the donor moiety There are amino acids having a donor in the side chain and those having an intramolecular hydrogen bond in the donor moiety
- two cyclic peptides containing each amino acid were synthesized, and the membrane permeability was compared by the improved method described in International Publication No. 2018/124162. ..
- the generality was confirmed by synthesizing and evaluating a plurality of combinations of the above two cyclic peptides.
- sequences of the two cyclic peptides to be compared were designed to minimize the influence of factors related to the membrane permeability other than the side chain partial structure having the donor.
- the amino acids to be compared were inserted so that they would be at the same position in the two cyclic peptides, and the number of amino acids constituting the cyclic part and linear part was made the same (resulting in the same total AA).
- the two cyclic peptides were arranged so that the number of donors in the side chain was the same (Thr and the like can be mentioned as examples of amino acids having a donor in the side chain in addition to the amino acid to be evaluated).
- the positions were designed to be the same.
- the number of N-alkyl groups in the amide group portion on the two main chains of the cyclic peptide was made uniform.
- N-alkyl groups eg NMe groups
- NH groups N-alkyl patterns
- amide groups eg, amide groups on two cyclic peptide main chains and Gly derivatives
- the positions and positions of cyclic amino acids such as Pro were designed to have the same conformation.
- the ClogP/totalAA value of the two peptides to be compared was adjusted to be between ⁇ 0.03 so that Trp was not included.
- the number of aromatic rings (Aromatic Ring Count: ARC) was made uniform.
- ClogP in this specification is a distribution coefficient calculated by a computer, and was calculated using Daylight Version 4.9 of Daylight Chemical Information Systems, Inc.
- A01 is a cyclic peptide consisting of 11 residues and forms a ring with 11 residues of amino acids.
- the cyclized portion is defined as the first amino acid, and Asp corresponds in the case of A01.
- the amino acid that constitutes the cyclic part next to Asp is defined as the second amino acid (MeLeu in the case of A01), and is repeated up to the third, fourth and N-terminals (the third MePhe, the fourth Ser(A01). iPen) and the cyclic part consists of 11 residues, so the N-terminus is the 11th D-Val).
- the C-term column means a functional group that condenses with the carboxylic acid site of the C-terminal amino acid, and pip means piperidine.
- Table 4 Table 6, Table 8, Table 10, Table 12, and Table 14, it is a peptide in which the N-terminal amino group and the side chain carboxylic acid of the amino acid in the cyclization part are cyclized by an amide bond.
- cyclic peptides A01 to B10 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- cyclic peptides C01 to D17 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- cyclic peptides E01 to F08 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- cyclic peptides G01 to H10 The structures of cyclic peptides G01 to H10 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- cyclic peptides I01 to J06 The structures of cyclic peptides I01 to J06 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- cyclic peptides AP01 to AP143 and AP200 to AP266 evaluated for membrane permeability are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- Example 3 Conformational analysis of side chain partial structure
- Example 3-1 Calculation Method of Low Energy Conformation of Side Chain Partial Structure
- the low energy conformation of the side chain partial structure uses a partial structure in which the side chain part of the amino acid is extracted from the side chain ⁇ position (carbon directly connected to the main chain). Calculated.
- Examples of Ser(EtOH), bAla(3R-MeOEtOH), bAla(2S-MeOEtOH), Phe(3-OMe-4-CONMe) are shown in the scheme below.
- Hyp(2-EtOH) pyrrolidine ring in the case of Hyp(2-EtOH)
- Hyp(2-EtOH) pyrrolidine ring in the case of Hyp(2-EtOH)
- Ser(EtOH), bAla(3R-MeOEtOH), bAla(2S-MeOEtOH), Hyp(2-EtOH) can be considered to have the same side chain partial structure.
- an amino acid of a type having a substituent on the amino group of an amino acid such as (Ph(3-OMe-4-CONMe)Et)Gly has a partial structure in which the side chain portion of the amino acid is extracted from the atom directly connected to the N atom.
- Low energy conformation was calculated by the following procedure.
- (1) Generation of initial structure The initial dihedral angles of rotatable single bonds were given as 0, 90, 180 and 270 degrees. The initial dihedral angle was given as 0,180 degrees for the amide bond and the bond between C and OH.
- the initial dihedral angles of 4 for each of the two single bonds and the initial dihedral angles of 2 for the bond between C and OH are taken into consideration.
- 32 initial conformations were generated.
- Structural optimization Using Gaussian09, structural optimization of all the conformations generated in (1) was performed. Structural optimization in pseudo water was performed by adding Polarizable Continuum Model (PCM) to B3LYP/6-31G(d) level.
- PCM Polarizable Continuum Model
- Example 3-2 Calculation Method of Potential Energy Surface 1
- potential energy surface (FIG. 3-1) in the following partial structure (eg, side chain partial structure of Ser(NMe-Aca))
- Low energy conformations were calculated by the above procedure.
- Energy calculation of each conformation (grid point)
- ⁇ 1 is 180° ⁇ 2
- ⁇ 3 is 0°
- 15°...24x24 576 as grid points are generated at 15° intervals up to 345°.
- ⁇ 1 is 0°
- 270° ⁇ 2 is 0°
- ⁇ 3 is 0° 180°
- ⁇ 2 180°
- 90°, 180°, 270° ⁇ 3 0°
- ⁇ 2 fixed at the initial value
- Structural optimization in pseudo water was performed by adding Polarizable Continuum Model (PCM) to B3LYP/6-31G(d) level.
- PCM Polarizable Continuum Model
- E( ⁇ 2) The lowest energy among E( ⁇ 2) was defined as E lowest ( ⁇ 2).
- the energies ( ⁇ E( ⁇ 2)) of 24 conformations in which ⁇ 2 was calculated at 15° intervals were calculated by the following formula.
- Example 3-3 Investigation of conformational distribution of crystal structures (FIGS. 3-2 and 4-2) using the results of X-ray structural analysis registered in CSD (The Cambridge Structural Database) The following queries were searched using CSD (The Cambridge Structural Database) 5.39+2 updates.
- amide NH group which is the donor moiety that forms the intramolecular hydrogen bond
- Fig. 3-1 membrane permeation
- intermolecular hydrogen bond that is advantageous for bond formation when interacting with a target. It was confirmed that the conformation can be set according to the surrounding environment, such as the conformation that forms the.
- Example 3-4 Low energy conformation calculation result of side chain partial structure
- the side chain partial structure is computationally shown to be a structure having an intramolecular hydrogen bond and easily in a conformation. It was Ser(EtOH), bAla(3R-MeOEtOH), bAla(2S-MeOEtOH), Hyp(2-EtOH): These three amino acids have the same side chain partial structure. The calculation result shows that the conformation having intramolecular hydrogen bond is the most stable conformation.
- Ser(1-CF3-EtOH) This amino acid has a racemic carbon moiety to which a CF 3 group is bonded, and therefore Ser (R-1-CF3-EtOH) which is an R-form and Ser (R-CF3-EtOH) which is an S-form
- the stable conformation was calculated for each (S-1-CF3-EtOH). In all the calculation results, the conformation having an intramolecular hydrogen bond was the most stable conformation. Only the most stable conformation is described in the calculation result column of Table 18.
- Ser (3-Me-5-Oxo-Odz) The calculation result shows that the conformation having an intramolecular hydrogen bond is 0.89 kJ/mol away from the most stable conformation.
- the column of the calculation result in the table describes the column that is 0.89 kJ/mol away from the most stable conformation.
- the side chain of an amino acid is a chain bonded to a carbon atom (eg, ⁇ , ⁇ , or ⁇ -carbon atom) contained in an amino acid. , And chains attached to the nitrogen atom.
- the side chain length of amino acids can be determined by the following method.
- the N-terminal of the amino acid unit is capped with an acetyl group and the C-terminal with a methylamino group, a conformation is generated by LowModeMD of molecular modeling software MOE (Chemical Computing Group), and the atom to which the side chain site is bonded ( In the case of a natural amino acid, it can be determined by measuring the distance from the ⁇ -carbon atom (C ⁇ carbon)) to the farthest atom on the same side chain (excluding hydrogen atom). As an example, the method for determining the length of the side chain of Phe or nBuGly is shown below.
- the amino acid side chain forms a ring with a part of the main chain structure, there are a plurality of atoms to which the side chain part is bonded.
- the length of the side chain is calculated by performing the same calculation as above for the atom to which each side chain is bonded and the atom of the farthest side chain (excluding hydrogen atom), and calculate the longest distance. Let's be long.
- Hyp(Et) the side chain distance obtained from the nitrogen atom is 6.04 ⁇
- the side chain distance obtained from C ⁇ carbon is 6.09 ⁇ .
- the distance is 6.09 ⁇ .
- the length of the amino acid side chain calculated by the above method is shown below.
- the calculated pKa of the side chain force ruboxi group of Asp is 4.3
- the calculated pKa of the side differential phenolic hydroxyl group of Tyr is 9.9
- 3-fluorotyrosine (Tyr(Tyr( 3-F)) had a calculated pKa for the laterally differential phenolic hydroxyl group of 8.7 and a calculated pKa for tetrazole of 3.7.
- the base the basic calculated pKa of the side chain guanidino group of Arg was 12.7
- the basic calculated pKa of the imidazolyl group of His was 7.6, and the basic calculated pKa of pyridine was 5.4. .
- Examplementation procedure 0.15 M KCl was added to about 100 mg of a test substance powder (about 1 mg) or 10 mM DMSO solution, and in the case of a basic compound, 0.5 M HCl was added until pH 2 was reached, and then 0.5 M KOH was added. Titration was performed to pH 12. In the case of an acidic compound, 0.5 M KOH was added until the pH reached 12, and then titration was performed with 0.5 M HCl until the pH reached 2. The titration operation was automatically performed by T3 manufactured by Sirius, and pKa was determined from the obtained titration curve using sirius T3 Refinement software.
- pKa was similarly determined using imipramine.HCl, propranolol.HCl, and warfarin as reference compounds, and it was confirmed that the pKa was within the test result range obtained by Sirius.
- the pKa of the amino acid side chain was determined by synthesizing a sequence in which the amino acid to be evaluated was introduced into the second residue (position X in the table below) of the peptide consisting of the three residues shown below.
- N-(((9H-fluoren-9-yl)methoxy)carbonyl)-N-methylglycine-2-chlorotrityl resin (compound pd11, Fmoc-MeGly-Trt(2-Cl)-resin) (14.4 mg) was placed in a reaction vessel, DMF (2 mL) was added, and the mixture was shaken for 1 hour. DBU (0.04 mL) was added to the reaction solution, shaken for 30 minutes, DMF (10 mL) was added, 1 mL was taken out, and further diluted with DMF until the solution amount became 12.5 mL.
- the absorbance (294 nm) of the obtained solution was measured (Shimadzu, UV-1600PC (cell length 1.0 cm) was used), and N-(((9H-fluoren-9-yl)methoxy)carbonyl)-N was measured.
- the loading amount of -methylglycine-2-chlorotrityl resin (compound pd11, Fmoc-MeGly-Trt(2-Cl)-resin) was calculated to be 0.789 mmol/g.
- the sequence containing Ser(Et-2-NMe2) which is an amino acid having a measured basic pKa of 9.1 and a calculated basic pKa of 8.9, has a range of 1.21 ⁇ ClogP/total aa ⁇ 1.36.
- MeAbu pip-4-F2
- MeAbu MeAbu
- Ser Et-2-Mor
- MeAbu pip-3-F2
- Abu Abu (5-Oxo-Odz) in the sequence including the membrane permeability. Examples of those achieving P app ⁇ 1.0 ⁇ 10 ⁇ 6 cm/sec are shown below (pd386-pd395, pd452-pd454, pd482-pd487).
- cyclic peptides pd50 to pd70 are shown below. (Note that in the structural formula of the cyclic peptide, the hydrogen atom bonded to the hetero atom may be omitted in order to clarify the structure.)
- the obtained residue was dissolved in 2,2,2-trifluoroethanol (10 mL), trimethylchlorosilane (745 ⁇ L, 5.83 mmol) was added, and the mixture was stirred at room temperature for 1 hr.
- the reaction mixture was concentrated under reduced pressure, the obtained residue was dissolved in 1,4-dioxane (5 mL) and 2N hydrochloric acid aqueous solution (10 mL), and the mixture was stirred at room temperature for 1 hr.
- the reaction solution was extracted twice with tert-butyl methyl ether, and the organic layer was concentrated under reduced pressure.
- the reaction mixture was concentrated under reduced pressure, dissolved in dichloromethane, washed with saturated aqueous sodium hydrogen carbonate solution, water, and saturated brine, and the obtained organic layer was dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure.
- the obtained residue was purified by normal phase silica gel column chromatography (petroleum ether/ethyl acetate) to obtain an intermediate (S)-4-(2-(4,4-difluoropiperidin-1-yl)ethyl). There was obtained (9H-fluoren-9-yl)methyl-5-oxooxazolidin-3-carboxylic acid (10.9 g, 82%).
- the obtained intermediate (6 g, 13.14 mmol) was dissolved in trifluoroacetic acid (65 mL) and dichloroethane (65 mL), triethylsilane (13.5 g, 116.1 mmol) was added at room temperature, and the temperature was 70° C. for 4 hours. It was stirred. The reaction solution was returned to room temperature and then concentrated under reduced pressure, and the obtained residue was dissolved in dichloromethane. The organic layer was washed with water, dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure.
- N′-Dicyclohexylcarbodiimide (2.756 g, 1.337 mmol) was added. After stirring at room temperature for 5 hours, the solid was removed by filtration. The filtrate was diluted with dichloromethane (DCM), washed with saturated brine, dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure.
- DCM dichloromethane
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Health & Medical Sciences (AREA)
- Life Sciences & Earth Sciences (AREA)
- Molecular Biology (AREA)
- Biochemistry (AREA)
- Medicinal Chemistry (AREA)
- General Health & Medical Sciences (AREA)
- Genetics & Genomics (AREA)
- Proteomics, Peptides & Aminoacids (AREA)
- Biophysics (AREA)
- General Chemical & Material Sciences (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Gastroenterology & Hepatology (AREA)
- Peptides Or Proteins (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Medicines That Contain Protein Lipid Enzymes And Other Medicines (AREA)
Abstract
Description
〔1〕2以上のアミノ酸が連結されたペプチド化合物であって、アミノ酸の少なくとも1つがその側鎖内で水素結合を形成し得る、ペプチド化合物。
〔2〕側鎖内で水素結合を形成し得るアミノ酸が、側鎖内で擬似4~7員環を形成し得る、〔1〕に記載のペプチド化合物。
〔3〕側鎖内で水素結合を形成し得るアミノ酸が、下記式Aで表される、〔1〕または〔2〕に記載のペプチド化合物:

R1は、水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、この場合、R2AおよびR2Bは、R2Aが水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、かつR2Bは、水素またはC1-C6アルキルであるか、あるいは、R2AおよびR2Bは、それらが結合している炭素原子と一緒になって4~6員の環を形成し、あるいは
R1は、R1が結合する窒素原子、R2A、およびR2Aが結合する炭素原子と一緒になって4~6員の複素環を形成し、該複素環は、式1もしくは式2で表される基、-OH、およびアルコキシ基からなる群より選択される、1つまたは複数の置換基を有していてもよく、この場合、R2Bは、水素またはC1-C6アルキルであり、
R3は、単結合、または-CHR4-であり、
R4は、水素、もしくはC1-C4アルキルであるか、または式1もしくは式2で表される基であり、
式1および式2は、それぞれ下記式:

*は、結合点を示し、
Q1およびQ2は、独立して、単結合、C1-C4アルキレン、または酸素原子を1つ含むC2-C4ヘテロアルキレンであり、
A1は、-O-、または-S-であり、
L1は、フッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
A2は、単結合、-O-、または-S-であり、
L2は、単結合であるか、またはフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xは、-OH、-NRZ1RZ2、もしくは-CONRZ1RZ2であるか、または1~3個のヘテロ原子を含有し、オキソもしくは1つまたは複数のハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり;
RZ1およびRZ2は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Yは、-OH、C1-C4アルキルスルホニルアミノ、-NRZ3RZ4、もしくは-CONRZ3RZ4であるか、または1~3個のヘテロ原子を含有し、オキソもしくはハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり、
RZ3およびRZ4は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Zは、水素またはC1-C4アルキルであり、
但し、A2が-O-、または-S-である場合、Zは水素ではなく、
ここで、式Aで表されるアミノ酸は、式1または式2で表される基のいずれかを少なくとも1つ含む。
〔4〕R1が、式1または式2で表される基であり、
R2AおよびR2Bが、独立して水素、もしくはC1-C6アルキルであるか、あるいは、R2AおよびR2Bは、それらが結合している炭素原子と一緒になって4~6員の環を形成し、
R3が、単結合、または-CHR4-であり、
R4が、水素、またはC1-C4アルキルである
〔3〕に記載のペプチド化合物。
〔5〕R1が、R1が結合する窒素原子、R2A、およびR2Aが結合する炭素原子と一緒になって4~6員の複素環を形成し、該複素環は、式1または式2で表される基を有しており、
R2Bが、水素またはC1-C6アルキルであり、
R3が、単結合、または-CHR4-であり、
R4が、水素、またはC1-C4アルキルである
〔3〕に記載のペプチド化合物。
〔6〕R1が、水素、もしくはC1-C6アルキルであり
R2Aが、式1もしくは式2で表される基であり、
R2Bは、水素またはC1-C6アルキルであり、
R3が、単結合、または-CHR4-であり、
R4が、水素、またはC1-C4アルキルである
〔3〕に記載のペプチド化合物。
〔7〕R1が、水素、もしくはC1-C6アルキルであり、
R2AおよびR2Bが、独立して水素、もしくはC1-C6アルキルであるか、あるいは、R2AおよびR2Bは、それらが結合している炭素原子と一緒になって4~6員の環を形成し、
R3が、-CHR4-であり、
R4が、式1もしくは式2で表される基である、
〔3〕に記載のペプチド化合物。
〔8〕Q1がC1-C4アルキレンであり、
A1が-O-または-S-であり、
L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xが-OHである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔9〕Q1がC1-C4アルキレンであり、
A1が-O-または-S-であり、
L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xが-CONRZ1RZ2であり、
RZ1がC1-C4アルキルであり、
RZ2が水素である、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔10〕Q1がC1-C4アルキレンであり、
A1が-O-または-S-であり、
L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xが1~3個のヘテロ原子を含有し、オキソもしくは1つまたは複数のハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔11〕Q1が単結合であり、
A1が-O-または-S-であり、
L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xが-OHである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔12〕Q1がCH2-である、〔3〕~〔11〕のいずれかに記載のペプチド化合物。
〔13〕A1が-O-である、〔3〕~〔12〕のいずれかに記載のペプチド化合物。
〔14〕L1が、-CH2-、-(CH2)2-、-(CH2)3-、-CH2CH(CH3)-、-CH2CH(CF3)-、-CH2C(CH3)2-、-(CH2)2CH(CH3)-、-CH2C(CH3)2CH2-、-(CH2)2C(CH3)2-、および-CH2CO-からなる群より選択される、〔3〕~〔13〕のいずれかに記載のペプチド化合物。
〔15〕Q2がC1-C4アルキレンであり、
L2が単結合であり、
Yが-CONRZ3RZ4であり、
A2が-O-、または-S-であり、
ZがC1-C4アルキルである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔16〕Q2がC1-C4アルキレンであり、
L2が単結合であり、
Yが-CONRZ3RZ4であり、
A2が単結合であり、
Zが水素である、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔17〕Q2がC1-C4アルキレンであり、
L2が単結合であり、
YがC1-C4アルキルスルホニルアミノカルボニルであり、
A2が-O-、または-S-であり、
ZがC1-C4アルキルである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔18〕Q2がC1-C4アルキレンであり、
L2が単結合であり、
Yが-OHであり、
A2が-O-、または-S-であり、
ZがC1-C4アルキルである、
〔3〕~〔7〕のいずれかに記載のペプチド化合物。
〔19〕Q2が-CH2-である、〔3〕~〔7〕、〔16〕~〔18〕のいずれかに記載のペプチド化合物。
〔20〕Yが、メチルアミノカルボニル、-CON(OH)Me、または-CONH(OH)である、〔3〕~〔7〕、〔15〕、および〔19〕のいずれかに記載のペプチド化合物。
〔21〕Yが、-CON(OH)Meである、〔3〕~〔7〕、〔16〕、および〔19〕のいずれかに記載のペプチド化合物。
〔22〕A2が-O-である、〔3〕~〔7〕、〔15〕、および〔17〕~〔21〕のいずれかに記載のペプチド化合物。
〔23〕Zがメチルである、〔3〕~〔7〕、〔15〕、〔17〕~〔22〕のいずれかに記載のペプチド化合物。
〔24〕式Aで表されるアミノ酸が以下からなる群:


〔25〕5~30のアミノ酸から構成される、〔1〕~〔24〕のいずれかに記載のペプチド化合物。
〔26〕環状である、〔1〕~〔25〕のいずれかに記載のペプチド化合物。
〔27〕2~15のアミノ酸から構成される環状部を含む、〔26〕に記載のペプチド化合物。
〔28〕2~30のN置換アミノ酸を含む、〔1〕~〔27〕のいずれかに記載のペプチド化合物。
〔29〕全アミノ酸数に対するN置換アミノ酸数の割合が30%以上である、〔1〕~〔28〕のいずれかに記載のペプチド化合物。
〔30〕4.0~18のClogPを有する、〔1〕~〔29〕のいずれかに記載のペプチド化合物。
〔31〕1.0x10⁻⁷cm/秒以上のPappを有する、〔1〕~〔30〕のいずれかに記載のペプチド化合物。
〔32〕〔1〕~〔31〕のいずれかに記載のペプチドおよび/または当該ペプチドをコードする核酸を含むライブラリ。
〔33〕〔3〕~〔23〕のいずれかに記載の式Aで表されるアミノ酸。
〔34〕以下からなる群:


〔35〕〔33〕または〔34〕に記載のアミノ酸に含まれるアミノ基、および/またはカルボキシル基が保護基で保護されている、保護アミノ酸。
〔36〕アミノ基の保護基が、Fmoc基、Boc基、Cbz基、Alloc基、ノシル基、ジニトロノシル基、t-Bu基、トリチル基、およびクミル基からなる群より選択される、および/または
カルボキシル基の保護基が、メチル基、アリル基、t-Bu基、トリチル基、クミル基、メトキシトリチル基、およびベンジル基からなる群より選択される、〔35〕に記載の保護アミノ酸。
本明細書において「アルキル」とは、脂肪族炭化水素から任意の水素原子を1個除いて誘導される1価の基であり、骨格中にヘテロ原子(炭素及び水素原子以外の原子をいう。)または不飽和の炭素-炭素結合を含有せず、水素及び炭素原子を含有するヒドロカルビルまたは炭化水素基構造の部分集合を有する。該アルキル基は直鎖状、又は分枝鎖状のものを含む。アルキル基としては、炭素原子数1~20(C1-C20、以下「Cp-Cq」とは炭素原子数がp~q個であることを意味する。)のアルキル基であり、好ましくはC1-C6アルキル基、C1-C5アルキル基、C1-C4アルキル基、C1-C3アルキル基などが挙げられる。アルキルとしては、具体的には、メチル、エチル、プロピル、ブチル、ペンチル、ヘキシル、イソプロピル、tert-ブチル基、sec-ブチル基、1-メチルプロピル基、1,1-ジメチルプロピル基、2,2-ジメチルプロピル、1,2-ジメチルプロピル、1,1,2-トリメチルプロピル、1,2,2-トリメチルプロピル、1,1,2,2-テトラメチルプロピル、1-メチルブチル、2-メチルブチル、3-メチルブチル、1,1-ジメチルブチル、1,2-ジメチルブチル、1,1-ジメチルブチル、1,2-ジメチルブチル、1,3-ジメチルブチル、2,2-ジメチルブチル、2,3-ジメチルブチル、3,3-ジメチルブチル、1-エチルブチル、2-エチルブチル、イソペンチル、ネオペンチル等が挙げられる。
ある局面において、本発明は、2以上のアミノ酸が連結されたペプチド化合物であって、アミノ酸の少なくとも1つ(具体的には、例えば、1つ、2つ、3つ、4つ、またはそれ以上)がその側鎖内で水素結合を形成し得るアミノ酸である、ペプチド化合物に関する。
本発明における「ペプチド化合物」には、2以上のアミノ酸が連結された、直鎖状または環状のペプチド化合物が含まれる。なお、環状ペプチド化合物は、「環状部を有するペプチド化合物」と同意義である。
(-C=O-OR)
(A)インドール骨格;
(B)2以上の芳香環による縮環構造;
(C)無置換のヒドロキシフェニル基。
このうち、環状部の側鎖に前記(A)、(C)を有しない環状ペプチド化合物が好ましく、環状部の側鎖に前記(B)、(C)を有しない環状ペプチド化合物がより好ましい。前記「2以上の芳香環による縮環構造」は、「縮環構造」であってもよい。
非限定の一態様において、本明細書におけるペプチド化合物、または環状ペプチド化合物は、高い膜透過性を達成するために、中性(例えばpH=7.0)で極度にイオン化される官能基を有しない環状ペプチド化合物であってよい。本明細書で用いるpKaは別段示さない限り実測値のpKaのことを指す。また、後述するADMETPredictorによって求めたpKa値を計算pKaと称する。本明細書で用いるBasic pKaは別段示さない限り実測値のBasic pKaのことを指す。また、後述するADMETPredictorによって求めたBasic pKa値をBasic計算pKaと称する。
本明細書における「膜透過係数(Papp)」は、特に断りがない限り、国際公開第2018/124162号に記載の測定方法(改良法)を用いて測定した値を意味する。
非限定の一態様において、本明細書におけるペプチド化合物は、良好な代謝安定性を有することが好ましい。良好な代謝安定性のためには、ペプチド化合物中に含まれるアミノ酸の数が8以上であることが好ましく、9以上がより好ましく、11以上が更に好ましい。また、一態様において、容易に酸化される可能性のあるチオエーテル結合を有しないことが好ましい。また、一態様において、酸化されやすく代謝安定性の妨げとなり得ることから、メチルチオ基を有しないことが好ましい。
非限定の一態様において、本発明のペプチド化合物は、その側鎖内で水素結合(即ち、分子内水素結合)を形成できるアミノ酸を少なくとも1つ、具体的には、例えば、1つ、2つ、3つ、4つ、またはそれ以上含む。


(i)R1が、式1または式2で表される基であり、R2Aが水素、またはC1-C6アルキル、好ましくは水素であり、R2Bが、水素またはC1-C6アルキル、好ましくは水素であり、R3が、単結合、または-CHR4-であり、R4が、水素、もしくはC1-C4アルキル、好ましくは水素である。
(ii)R1が水素またはC1-C6アルキルであり、R2Aが式1または式2で表される基であり、R2Bが水素またはC1-C6アルキル、好ましくは水素であり、R3が、単結合、または-CHR4-、好ましくは単結合であり、R4が、水素、もしくはC1-C4アルキル、好ましくは水素である。
(iii)R1が水素またはC1-C6アルキルであり、R2Aが水素、またはC1-C6アルキル、好ましくは水素であり、R2Bが水素、またはC1-C6アルキル、好ましくは水素であり、R3が-CHR4-であり、R4が式1または式2で表される基である。
(iv) R1、R1が結合する窒素原子、R2A、およびR2Aが結合する炭素原子が一緒になって、4~6員の複素環(好ましくはピロリジン環)を形成し、該複素環が式1もしくは式2で表される基を有し、R2Bが、水素またはC1-C6アルキル、好ましくは水素であり、R3が、単結合、または-CHR4-、好ましくは単結合であり、R4が、水素、もしくはC1-C4アルキル、好ましくは水素である。


(i) Q1がC1-C4アルキレン、好ましくは、-CH2-であり、A1が-O-または-S-、好ましくは-O-であり、L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、Xが-OHである。
(ii) Q1がC1-C4アルキレン、好ましくは、-CH2-であり、A1が-O-または-S-、好ましくは-O-であり、L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは、-CH2-であり、Xが-NRZ1RZ2であり、RZ1がC1-C4アルキルであり、RZ2が水素である。
(iii) Q1がC1-C4アルキレン、好ましくは、-CH2-であり、A1が-O-または-S-、好ましくは-O-であり、L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは、-CH2-であり、Xが-CONRZ1RZ2であり、RZ1がC1-C4アルキルであり、RZ2が水素である。
(iv) Q1がC1-C4アルキレン、好ましくは、-CH2-であり、A1が-O-または-S-、好ましくは-O-であり、L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは、-CH2-であり、Xが1~3個のヘテロ原子を含有し、オキソもしくは1つまたは複数のハロゲン(好ましくはフッ素)によって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルである。
(v) Q1が単結合であり、A1が-O-または-S-、好ましくは-O-であり、L1が、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、Xが-OHである。
(i) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2がフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは-CO-であり、Yが-NRZ3RZ4、好ましくはメチルアミノ、-N(OH)Me、または-NH(OH)であり、A2が-O-、または-S-、好ましくは-O-であり、ZがC1-C4アルキル、好ましくはメチルである。
(ii) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2が単結合であり、Yが-CONRZ3RZ4、好ましくはメチルアミノカルボニルまたは-CON(OH)Me、または-CONH(OH)であり、A2が-O-、または-S-、好ましくは-O-であり、ZがC1-C4アルキル、好ましくはメチルである。
(iii) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2がフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは-CO-であり、Yが-NRZ3RZ4、好ましくは-N(OH)Meであり、A2が単結合であり、Zが水素である。
(iv) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2が単結合であり、Yが-CONRZ3RZ4、好ましくは-CON(OH)Meであり、A2が単結合であり、Zが水素である。
(v) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2がフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレン、好ましくは-CO-であり、YがC1-C4アルキルスルホニルアミノ、好ましくはメチルスルホニルアミノであり、A2が-O-、または-S-、好ましくは-O-であり、ZがC1-C4アルキル、好ましくはメチルである。
(vi) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2が単結合であり、YがC1-C4アルキルスルホニルアミノカルボニル、好ましくはメチルスルホニルアミノカルボニルであり、A2が-O-、または-S-、好ましくは-O-であり、ZがC1-C4アルキル、好ましくはメチルである。
(vii) Q2がC1-C4アルキレン、好ましくは、-CH2-であり、L2が単結合であり、Yが-OHであり、A2が-O-、または-S-、好ましくは-O-であり、ZがC1-C4アルキル、好ましくはメチルである。


次に本発明の側鎖内で水素結合を形成し得るアミノ酸、およびペプチド化合物の一般的な製造方法について説明する。
側鎖に分子内水素結合を有するアミノ酸の製造において、側鎖にエーテル結合を有するアミノ酸に関しては、以下の2つの合成法により製造できる。
アミノ基とカルボン酸基が保護されたアジリジンを有する化合物と対応する適切な保護基を有するアルコールをLewis酸存在下反応させ、開環反応を行った後、不要な保護基の脱保護を行う。下記一般式中、P1としてはCbz基、Fmoc基、Alloc基、ノシル基(Ns基)などを、P2としてはMe基、アリル基、ベンジル基などを用いることが出来る。開環反応に用いるLewis酸としては触媒量のBF3・OEt2が好ましく例示される。溶媒はジクロロメタンが好ましく例示されるが、試薬として使用するアルコール(ROH)を溶媒として用いることも可能である。R上に反応に関与する可能性のある官能基が含まれる場合、該官能基をLewis酸存在下において安定な保護基で保護することが好ましい。例えばRが水酸基を含む場合はベンジル基などで該水酸基を、2級アミノ基を含む場合はAlloc基などで該アミノ基を保護することが好ましい。開環反応によりアミノ酸の基本骨格を構築した後は保護基の適切な変換を行うことで所望のアミノ酸(P3がCbz基、Fmoc基、Boc基、Alloc基、ノシル基(Ns基)などで保護されたアミノ酸あるいはP3が水素である無保護のアミノ酸)に変換できる。

側鎖に水酸基を有する、アミノ基が保護されたアミノ酸と2―ブロモ―N―メチルアセトアミドのようなαハロカルボニル化合物あるいはブロモアセトニトリルとのアルキル化反応を行った後、適切な官能基変換により下記スキームに示す製造を実施することができる。下記一般式中、P1としてはCbz基、Boc基、Trt基などの塩基性条件下で安定な保護基が好ましい。αハロカルボニル化合物としては、RaがNMeやNtBuなどであるものが好ましく例示される。X1はハロゲンであり、ブロモ基が好ましい。アルキル化反応において用いられる塩基はNaH、NaOtBu、NaOt-Pentなどが例示され、側鎖に水酸基を有する、アミノ基が保護されたアミノ酸に対し、2当量以上用いることで選択的に該アミノ酸の水酸基部位をアルキル化することができる。用いる溶媒としてはDMF、DMIなどが好ましく例示される。ブロモアセトニトリルを用いたアルキル化の後、得られたニトリル基を元にヘテロ環を構築することでアミノ酸基本骨格を構築できる。下記スキームには生じたニトリル基にヒドロキシルアミンを作用させた後、CDIで環化したオキサジアゾロン環を構築したものを例示している。アミノ酸の基本骨格を構築した後は保護基の適切な変換を行うことで所望のアミノ酸(P3がCbz基、Fmoc基、Boc基、Alloc基、ノシル基(Ns基)などで保護されたアミノ酸あるいはP3が水素である無保護のアミノ酸)に変換できる。

フェニルアラニン誘導体はJackson-Negishi coupling (Journal of Organic Chemistry, 2010, 75, 245)に記載の方法や、相関移動触媒を用いたグリシンtert-ブチルエステルのベンゾフェノンイミンの不斉ベンジル化(J. Am. Chem. Soc. 1999, 121, 6519)などを用いて製造することができる。また、下記一般式に示すようなN-ヒドロキシフタルイミドエステル(NHPIエステル)と芳香族ヨウ素化合物あるいは芳香族ブロモ化合物を原料に用いた脱炭酸をともなうクロスカップリング反応を行うことでも製造することができる。下記一般式中、nは炭素原子の数を表しており、特に限定はされないが、n=1、もしくは2が好ましい。P1としてはクロスカップリング条件下で安定な基が用いられ、好ましくはFmoc基が例示される。R4-X1は芳香族ハロゲン化合物であり、R4が芳香族部位を表している。R4にクロスカップリング反応条件下で変換されうる官能基が含まれる場合、クロスカップリング反応条件下において安定な保護基で保護することが好ましい。例えばR4に水酸基が含まれる場合、水酸基をテトラヒドロピラニル(THP)基などで保護することが好ましい。X1はハロゲンであり、好ましくはブロモ(Br)基、ヨード(I)基が例示される。クロスカップリング反応に用いる触媒としてはニッケルと亜鉛を触媒として用いるのが好ましく、触媒の配位子としては4,4’-ジ-tert-ブチル-2,2’-ビピリジル(dtbbpy)を用いるのが好ましい。溶媒としてはジメチルアセトアミドを用いるのが好ましい。脱炭酸をともなうクロスカップリング反応によりアミノ酸の基本骨格を構築した後は保護基の適切な変換を行うことで所望のアミノ酸(P3がCbz基、Fmoc基、Boc基、Alloc基、ノシル基(Ns基)などで保護されたアミノ酸あるいはP3が水素である無保護のアミノ酸)に変換できる。


アーント・アイシュタート(Arndt-Eistert)合成(人名反応に学ぶ有機合成戦略 、p18)を用いることで対応するβ―アミノ酸を製造することができる。

β―アミノ酸は、対応する官能基を有するエポキシドに対しアンモニアを用いた開環反応を行い、得られる化合物のアミン部位に保護基を導入し、次いで生じる2級アルコールを脱離基に変換した後、求核置換反応による立体反転を伴ったニトリル基への変換を行い、これに続くカルボン酸への変換を行うことで製造できる。下記一般式中、Rに反応に関与する官能基が含まれる場合、適切な保護基で保護することが好ましく、例えばRに水酸基が含まれる場合、該水酸基をTrt基で保護することが好ましい。開環反応後に得られるアミノ基の保護基であるP1としてはCbz基、Boc基、Alloc基などが好ましく例示される。生じた2級アルコールは脱離基X2へと変換され(X2としては好ましくはメシル基(OMs基)などが例示される)。得られた脱離基をシアノヒドリン(CN-)による求核置換反応により立体反転を伴いながらニトリル基へと誘導し、カルボン酸へと変換することでアミノ酸の基本骨格を構築できる。ニトリル基からカルボン酸への変換は例えば酸性条件下における加水分解が例示される。アミノ酸の基本骨格を構築した後は保護基の適切な架けかえを行うことで所望のアミノ酸(P3がCbz基、Fmoc基、Boc基、Alloc基、ノシル基(Ns基)などで保護されたアミノ酸あるいはP3が水素である無保護のアミノ酸)に変換できる。

水酸基を有し、かつアミノ基が保護された環状アミノ酸とハロゲン化合物とを、塩基を用いたアルキル化反応に供した後、適切な官能基の変換を行うことにより下記スキームに示すとおりに環状アミノ酸を製造することができる。用いる原料の環状アミノ酸としては4~8員環のものが好ましく、下記スキームでは5員環のものを例示した。環状部分に存在する水酸基の位置は特に限定されないが、一般式にあるようにアミノ酸のN原子から2原子以上離れた位置にあることが好ましい。下記一般式中、P1としてはCbz基、Boc基、Alloc基、ノシル基(Ns基)などを用いることが出来る。アルキル化反応に用いる塩基としては水素化ナトリウム(NaH)が好ましく例示され、溶媒はジメチルホルムアミドが好ましく例示される。アルキル化反応において、ハロゲン化合物(R-X)がR上に該反応に関与する可能性のある官能基を有する場合、その官能基を塩基性条件下において安定な保護基で保護することが好ましい。例えばRが水酸基を含む場合はテトラヒドロピラニル(THP)基、ベンジル(Bn)基、t-ブチルジメチルシリル(TBDMS)基などのシリルエーテルなどで該水酸基を保護することが好ましい。またX1はハロゲンであり、クロロ(Cl)基、ブロモ(Br)基、ヨード(I)基が例示される。アルキル化反応によりアミノ酸の基本骨格を構築した後は保護基の適切な変換を行うことで所望のアミノ酸(P3がCbz基、Fmoc基、Boc基、Alloc基、ノシル基(Ns基)などで保護されたアミノ酸あるいはP3が水素である無保護のアミノ酸)に変換できる。

上記のアミノ酸は、下記スキームに沿って、酸性条件下、パラホルムアルデヒドを用いた環化反応を行い、続いて得られた環化体を酸性条件下で開環しながら生じるイミンを還元することで、N-メチル化を行うことができる。

ペプチド化合物の化学合成方法
本明細書におけるペプチド化合物、または環状ペプチド化合物の化学合成方法としては、液相合成法、Fmoc合成やBoc合成等を用いた固相合成法、及びこれらの組み合わせ等が例示される。Fmoc合成では、主鎖アミノ基がFmoc基により保護され、側鎖官能基が必要に応じてピペリジンやt-Bu基、THP基、Trt基などの塩基性で切断されない保護基で保護され、主鎖カルボン酸を保護しないアミノ酸を基本単位とする。基本単位としては、その他、Fmoc保護されたアミノ基とカルボキシル基を有する組み合わせであれば、特に限定されない。例えばジペプチドを基本単位としてもよい。N末端に配置させる基本単位は、Fmocアミノ酸以外であってもよい。例えば、Bocアミノ酸であってもよく、アミノ基を有しないカルボン酸類縁体であってもよい。主鎖カルボキシル基あるいは主鎖カルボキシル基が適切な保護基で保護された側鎖にカルボキシル基を有するアミノ酸の側鎖カルボキシル基を固相担体の官能基との化学反応により固相に担持させる。続いてピペリジンやDBUなどの塩基によりFmoc基を脱保護し、新たに生じたアミノ基と続いて添加される基本単位のカルボキシル基を有する保護アミノ酸とを縮合反応させることで、ペプチド結合を生成させる。縮合反応では、カルボキシル基の活性化剤として、DICとHOBtの組み合わせ、DICとHOAtの組み合わせ、HATUとDIPEAとの組み合わせなど様々な組み合わせが可能である。脱Fmoc基とそれに続くペプチド結合生成反応の繰り返しにより、望みのペプチド配列を生成させることができる。望みの配列が得られた後、固相からの切り出しと必要に応じて導入した側鎖官能基の保護基の脱保護が行われる。また、固相からの切り出しを行う前にペプチドの構造変換や環化を行うことも可能である。固相からの切り出しと、脱保護は同一の条件、例えば90:10のTFA/H2Oで行っても良いし、必要に応じて別々の条件で脱保護しても良い。固相からの切り出しは、1% TFAなどの弱酸での切り出しが可能なものもあれば、保護基としてPdなどを利用して、両者の化学反応の直行性を利用することが可能である。これらの工程の間もしくは最後に、環化などの工程を実施することもできる。例えば、側鎖カルボン酸とN末端主鎖のアミノ基とを縮合させることや、側鎖アミノ基とC末端主鎖のカルボン酸を縮合させることができる。この際、C末端側のカルボン酸と環化させる側鎖カルボン酸の間、もしくはN末端側の主鎖アミノ基またはヒドロキシ基と環化させる側鎖アミノ基の間で反応直行性が必要であり、前述のとおり保護基の直行性を考慮して保護基の選択が行われる。このようにして得られた反応物を逆相カラムや、分子ふるいカラムなどで精製することができる。これらの詳細は、例えばメルク株式会社が平成14年5月1日に発行した固相合成ハンドブックなどに記載されている。固相合成用樹脂は市販されているものを使うことができ、CTC樹脂、Wang樹脂、またはSASRIN樹脂が例示される。
本明細書におけるペプチド化合物の翻訳合成方法としては、無細胞翻訳系を用いて合成する方法が例示され、同方法には再構成無細胞翻訳系による合成法が含まれる。排除したい因子を除くことができる点で、再構成無細胞翻訳系を利用するのが好ましい。
(i) 本明細書におけるアミノ酸を少なくとも1種、2種以上、又は3種以上結合したtRNAを用意する工程;
(ii) 前記tRNAのアンチコドンに対応するコドンを少なくとも1つ含む核酸を無細胞翻訳系で翻訳し、前記ペプチド化合物を得る工程。
非限定の一態様において、本明細書における翻訳合成方法は、環状ペプチド化合物の翻訳合成方法であってよい。
本明細書において「無細胞翻訳系」とは、細胞より抽出したリボソームの他、翻訳に関与する蛋白因子群、tRNA、アミノ酸、ATPなどのエネルギー源、その再生系を組み合わせたもので、mRNAを蛋白質に翻訳することが出来るものであれば限定されない。また、翻訳合成反応が進行している間の系も、本明細書における「無細胞翻訳系」に含まれてよい。本明細書における無細胞翻訳系は、ペプチドを翻訳する際の鋳型となる核酸を含んでいてよく、これら以外にも、開始因子、伸長因子、解離因子、アミノアシルtRNA合成酵素などを含むことができる。これらの因子は、種々の細胞の抽出液から精製することによって得ることができる。因子を精製するための細胞は、例えば原核細胞、または真核細胞を挙げることができる。原核細胞としては、大腸菌細胞、高度好熱菌細胞、または枯草菌細胞を挙げることができる。真核細胞としては、酵母細胞、小麦胚芽、ウサギ網状赤血球、植物細胞、昆虫細胞、または動物細胞を材料にしたものが知られている。またtRNAやアミノアシルtRNA合成酵素(ARS)は天然に存在するものに加え、人工tRNAや非天然アミノ酸を認識する人工アミノアシルtRNA合成酵素を用いることもできる。人工tRNAや人工アミノアシルtRNA合成酵素を用いることにより部位特異的に非天然アミノ酸を導入したペプチドを合成することができる。なお、必要に応じて、無細胞翻訳系にT7 RNA polymeraseなどのRNAポリメラーゼを加えることで、鋳型DNAから転写させることもできる。
(i) 本明細書におけるアミノ酸を少なくとも1種、2種以上、又は3種以上結合したtRNA;
(ii) 前記ペプチド化合物をコードする核酸、
ここで該核酸は前記tRNAのアンチコドンに対応するコドンを少なくとも1つ含んでよい。
非天然アミノ酸のペプチドへの翻訳導入には、直交性を有し効率よくリボソームに取り込まれるtRNA((i)Biochemistry. 2003;42:9598-608.Adaptation of an orthogonal archaeal leucyl-tRNA and synthetase pair for four-base, amber, and opal suppression. Anderson JC, Schultz PG., (ii)Chem Biol. 2003;10:1077-84.Using a solid-phase ribozyme aminoacylation system to reprogram the genetic code.Murakami H, Kourouklis D, Suga H.)のアミノアシル化が必要である。tRNAをアミノアシル化する方法として以下の5つの方法を用いることができる。
非限定の一態様において、ペプチド化合物の環状部は、直鎖状のペプチド化合物を環化反応に供することにより形成される。

非限定の一態様において、本開示におけるペプチド化合物、又はこれらをコードする核酸のライブラリは、それぞれ、本明細書に記載のペプチド化合物、又はこれらをコードする核酸から実質的になるライブラリであってよい。
本明細書におけるライブラリには、本明細書におけるペプチド化合物を含むライブラリ、及び本明細書におけるペプチド化合物をコードする核酸を含むライブラリが含まれる。本明細書におけるライブラリには、本明細書におけるペプチド化合物のライブラリ、ペプチド化合物-核酸複合体のライブラリが含まれ、中でも環状ペプチド化合物、又は環状ペプチド-核酸複合体のライブラリが好ましく、特に環状ペプチド-mRNA複合体のライブラリが好ましい。ライブラリとしては、ディスプレイライブラリが好ましい。ディスプレイライブラリとしては、ディスプレイを利用したライブラリが例示され、中でもmRNAディスプレイライブラリ、DNAディスプレイライブラリ、リボソームディスプレイライブラリが好ましく、mRNAディスプレイライブラリがより好ましい。
ディスプレイライブラリは表現型であるペプチドとその遺伝子型であるペプチドをコードするRNAもしくはDNAが対応付けられているライブラリである。このライブラリを使用して、標的分子に特異的に結合できるペプチド化合物を同定することができる。例えば、所望の固定した標的にライブラリを接触させ、標的に結合しない分子を洗い流すことで、標的に結合するペプチドの濃縮が可能である(パニング法)。このような過程を経て選択されたペプチドに対応付けられている遺伝子情報を解析することで標的に結合したペプチドの配列を明らかにすることができる。例えば、アミノアシルtRNAのアナログである抗生物質ピューロマイシンがリボソームによるmRNA翻訳伸長中の蛋白質に非特異的に連結されることを利用する方法がmRNAディスプレイ(Proc Natl Acad Sci USA. 1997;94:12297-302. RNA-peptide fusions for the in vitro selection of peptides and proteins. Roberts RW, Szostak JW.)ないしはin vitro virus(FEBS Lett. 1997;414:405-8. In vitro virus: bonding of mRNA bearing puromycin at the 3'-terminal end to the C-terminal end of its encoded protein on the ribosome in vitro. Nemoto N, Miyamoto-Sato E, Husimi Y, Yanagawa H.)として報告されている。
本発明における「核酸」には、デオキシリボ核酸(DNA)、リボ核酸(RNA)あるいは、人工塩基を有するヌクレオチド誘導体を含むこともできる。また、ペプチド核酸(PNA)を含むこともできる。本発明の核酸は、目的とする遺伝情報が保持される限り、これらの核酸のいずれか、あるいは混成とすることもできる。すなわち、DNA-RNAのハイブリッドヌクレオチドやDNAとRNAのような異なる核酸が一本鎖に連結されたキメラ核酸も本発明における核酸に含まれる。
非限定の一態様において、本開示におけるライブラリの製造は、上述した本開示におけるペプチド化合物の製造方法に準じて行うことができ、適宜公知の方法と組み合わせることができる。一態様において、上述した本開示における無細胞翻訳系を使用して本開示におけるペプチド化合物のライブラリを製造できる。すなわち、本開示におけるライブラリの製造方法には、本開示における無細胞翻訳系を使用してペプチド化合物を合成する工程を含んでよい。一態様において、本開示における無細胞翻訳系において記述した、例示、好ましい範囲、態様が、本開示におけるライブラリの製造方法においてもそのまま適用できる。
(i) 本明細書に記載されたアミノ酸からなる群より選択される少なくとも1種、2種以上、又は3種以上のアミノ酸が結合したtRNA;
(ii) 前記ペプチド化合物ライブラリをコードする核酸ライブラリ、
ここで該核酸ライブラリには前記tRNAのアンチコドンに対応するコドンを少なくとも1つ含む核酸が含まれてよい。


LC/MSの分析条件は、下記のとおりである。


WO2013/100132に記載のFmoc法によるペプチド合成法に従い、下記の基本ルートでペプチドの伸長を行った。すなわち、1)Asp側鎖のカルボン酸を2-クロロトリチルレジンに担持させたものの、AspのN末からのFmoc法によるペプチド伸長反応、2)2-クロロトリチルレジンからのペプチドの切り出し過程、3)切り出し過程によって2-クロロトリチルレジンから外れて生じたAsp側鎖のカルボン酸と、ペプチド鎖N末端(三角ユニット)のアミノ基との縮合によるアミド環化、4)ペプチド鎖に含む側鎖官能基の保護基の脱保護、5)preparativeHPLCによる化合物の精製、の5段階の工程である。また、側鎖に酸性官能基を有するアミノ酸を含むペプチド化合物においては、脱保護剤の残留に起因するペプチドの過剰伸長を抑えるため、ペプチド伸長反応時のレジン洗浄に0.08 Mトリエチルアミン塩酸塩のジクロロメタン溶液を用いることもできる。本実施例において、特に記述がない限り、この基本ルートをもとにペプチド化合物の合成をおこなった。

本明細書内に記載するペプチド合成において、ペプチド合成機による合成には、以下のFmoc-アミノ酸を用いた。側鎖に反応性の官能基を有するアミノ酸の場合、適切な保護基で保護されているものを用いた。なおアミノ酸の略語は下記表3に記載した。
Fmoc-D-Val-OH、Fmoc-MeLeu-OH、Fmoc-MePhe-OH、Fmoc-MeGly-OH、Fmoc-gMeAbu-OH(Fmoc-g-MeAbu-OHと記載する場合もある)、Fmoc-Leu-OH、Fmoc-Ile-OH、Fmoc-MeAla-OH、Fmoc-MeVal-OH、Fmoc-D-Abu-OH、Fmoc-Val-OH、Fmoc―MeAbu-OH、Fmoc-MeIle-OH、Fmoc-D-Leu-OH、Fmoc-Ser(tBu)-OH、Fmoc-D-Ala-OH、Fmoc-Tyr(Clt)-OH、Fmoc-Phe-OH、Fmoc-Phe(4-CF3)-OH、Fmoc-Ala-OH、Fmoc-Hph-OH、Fmoc-Gly-OH、Fmoc-b-MeAla-OH、Fmoc-Ala(3-Pyr)-OH、Fmoc-D-MeAla-OH、Fmoc-Phe{#(CH2)2}-OH、Fmoc-Ala(Thz)-OH、Fmoc-D-Pro-OH、Fmoc-Pro-OH、Fmoc-Ser(Bn)-OHなどは渡辺化学もしくはChempep社もしくはChem-Impex社もしくはBachem社から購入した。またFmoc-nPrGly-OH、Fmoc-MePhe(3-Cl)-OH、Fmoc-MeAla(4-Thz)-OHなどはWO2013/100132に記載の方法にて合成した。また、Fmoc-Thr(THP)-OH(化合物aa01)、Fmoc-nBuGly-OH(化合物aa02)、Fmoc-Ser(iPen)-OH(化合物aa04)、Fmoc-MeHph-OH(化合物aa05)、Fmoc-Ser(3-F-5-Me-Pyr)-OH(化合物aa12)、Fmoc-Ser(Ph-3-Cl)-OH(化合物aa16)、Fmoc-Nle(6-OTHP)-OH(化合物aa17)、Fmoc-Ser(EtOTHP)-OH(化合物aa22)、Fmoc-MeSer(EtOTHP)-OH(化合物aa27)、Fmoc-Ser(S-2-PrOTHP)-OH(化合物aa34)、Fmoc-MeSer(S-2-PrOTHP)-OH(化合物aa39)、Fmoc-Ser(R-2-PrOTHP)-OH(化合物aa46)、Fmoc-MeSer(R-2-PrOTHP)-OH(化合物aa51)、Fmoc-Ser(tBuOTHP)-OH(化合物aa54)、Fmoc-MeSer(tBuOTHP)-OH(化合物aa55)、Fmoc-Ser(2-Me-2-BuOTHP)-OH(化合物aa62)、Fmoc-MeSer(2-Me-2-BuOTHP)-OH(化合物aa64)、Fmoc-Hnl(7-F3-6-OH)-OH(化合物aa68)、Fmoc-Ser(1-CF3-EtOH)-OH(化合物aa70)、Fmoc-MeSer(1-CF3-EtOH)-OH(化合物aa71)、Fmoc-Ser(1-CF3-EtOTHP)-OH(化合物aa72)、Fmoc-Gln(Me)-OH(化合物aa75)、Fmoc-Ser(NtBu-Aca)-OH(化合物aa78)、Fmoc-MeSer(NtBu-Aca)-OH(化合物aa79)、Fmoc-Ser(NMe-Aca)-OH(化合物aa81)、Fmoc-Ser(nPrOTHP)-OH(化合物aa85)、Fmoc-Ser(2-Me2-PrOTHP)-OH(化合物aa89)、Fmoc-Ser(S-2-BuOTHP)-OH(化合物aa95)、Fmoc-Ser(R-2-BuOTHP)-OH(化合物aa101)、Fmoc-Phe(4-OClt-3-OMe)-OH(化合物aa104)、Tyr(3-OMe)の側鎖フェノール部位がクロロトリチル基で保護されたもの)、Fmoc-Ser(3-Me-5-Oxo-Odz)-OH(化合物aa109)、Fmoc-bAla(3R-MeOEtOTHP)-OH(化合物aa111)、Fmoc-bAla(2S-MeOEtOTHP)-OH(化合物aa117)、Fmoc-Nva(2R-4-F2-Pyrro(N-Alloc))-OH(化合物aa121)、Fmoc-Ser(S-4-F2-Pyrro(N-Alloc)-Me)-OH(化合物aa126)、Fmoc-Phe(3-OMe-4-CONMe)-OH(化合物aa127)、Fmoc-Phe(4-OMe-3-CONMe)-OH(化合物aa128)、Fmoc-Phe(3-OMe-4-CONHMs)-OH(化合物aa129)、Fmoc-Hyp(2-EtOTHP)-OH(化合物aa133)、Fmoc-(Ph(3-OMe-4-CONMe)Et)Gly-OH(化合物134)、Fmoc-Hph(3-OMe-4-CONMe)-OH(化合物aa135)、Fmoc-D-Hph(3-OMe-4-CONMe)-OH(化合物aa136)、Fmoc-MeHph(3-OMe-4-CONMe)-OH(化合物aa137)、Fmoc-Phe(4-CONMeOTHP)-OH(化合物aa139)、Fmoc-Phe(3-OMe-4-CONHOTHP)-OH(化合物aa140)、Fmoc-Ser(S-(Alloc)Mor-3-Me)-OH(化合物aa141)などは以下のとおり合成した。
側鎖に3級アルコールを有するアミノ酸(例えば、Ser(tBuOH)、MeSer(tBuOH)、Ser(2-Me-2-BuOH)、MeSer(2-Me-2-BuOH)など)を含む配列に関しては、側鎖の3級アルコール部位をTHPで保護しないFmocアミノ酸を用いても問題無くペプチドは伸長した(例えば、Fmoc-Ser(tBuOH)-OH(化合物aa53)、Fmoc-Ser(2-Me-2-BuOH)-OH(化合物aa61)などを用いた)。
側鎖に2級アルコールであり、かつアルコールが結合している炭素にCF3基が結合している構造を持つアミノ酸(例えば、Hnl(7-F3-6-OH)、Ser(1-CF3-EtOH)など)を含む配列に関しては、側鎖の2級アルコール部位をTHPで保護しないFmocアミノ酸を用いても問題無くペプチドは伸長した(例えば、Fmoc-Hnl(7-F3-6-OH)-OH(化合物aa68)、Fmoc-Ser(1-CF3-EtOH)-OH(化合物aa70)などを用いた)。


(2S,3R)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブタン酸(化合物aa01、Fmoc-Thr(THP)-OH)の合成

得られた粗生成物のうち、4.65 gをテトラヒドロフラン(THF)(30 mL)に溶解させ、次いでpH8.0に調製した1.0Mリン酸緩衝液(30 mL)を加えた。この混合物を50度で4時間攪拌した。25度まで冷却した後、酢酸エチル(30 mL)を加え、有機層と水層を分離した。水層に酢酸エチル(30 mL)を加えて抽出を行った後、得られた全ての有機層を混合し、飽和塩化ナトリウム水溶液(30 mL)で2度洗浄した。有機層を硫酸ナトリウムで乾燥させ、溶媒を減圧下留去し、さらにポンプにて減圧下、25度で30分乾燥させた。
得られた残渣をジエチルエーテル(50 mL)に溶解させ、次いでヘプタン(50 mL)を加えた。制御した減圧下(~100 hPa)、ジエチルエーテルのみを留去し、得られた混合物をろ過して固体を得た。このヘプタンでの洗浄操作を2度繰り返した。得られた固体をポンプにて減圧下、25度で2時間乾燥させ、Fmoc-Thr(THP)-OHのナトリウム塩(2.80 g、6.26 mmol)を得た。
得られた全量のFmoc-Thr(THP)-OHのナトリウム塩に酢酸エチル(50 mL)とpH2.1の0.05Mリン酸水溶液(140 mL)を加えて、25度にて5分間攪拌した後、有機層と水層を分離した。水層に酢酸エチル(50 mL)を加えて抽出した後、得られた全ての有機層を混合し、飽和塩化ナトリウム水溶液(50 mL)にて2度洗浄した。有機層を硫酸ナトリウムにて乾燥させ、減圧下溶媒を留去した。残渣をポンプにて減圧下、25度で2時間乾燥させた後、得られた固体をt-ブチルメチルエーテル(TBME)(50 mL)に溶解させ、溶媒を減圧下留去した。さらにポンプにて減圧下、25度で1時間乾燥させることで、(2S,3R)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブタン酸(化合物aa01、Fmoc-Thr(THP)-OH、2.70 g、30 mol%のt-ブチルメチルエーテル(TBME)が残留)をTHP保護上の不斉炭素に由来するジアステレオマーとして得た。得られたFmoc-Thr(THP)-OHは-25度の冷凍庫にて保存した。
LCMS(ESI)m/z=424.2(M-H)-
保持時間:0.84分、0.85分(分析条件SQDFA05)

上述の方法で得たtert-ブチルN-(tert-ブトキシカルボニル)-N-ブチルグリシナート(Boc-nBuGly-OtBu)(260 g、904.68 mmol)の1,4-ジオキサン溶液(1000 mL)に濃塩酸(1000 mL)を0度にて滴下しながら加えた。反応液を室温に昇温した後、16時間室温にて撹拌した。反応液を濃縮することでブチルグリシン(H-nBuGly-OH)(200 g)の塩酸塩を粗生成物として得た。
粗生成物であるブチルグリシン(H-nBuGly-OH)(60 g)、炭酸カリウム(188.9 g、1.37 mol)、水/1,4-ジオキサン(1:1)(3000 mL)の混合物を室温にて30分間撹拌した後、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(183.2 g、543.09 mmol)を少しずつ室温にて加えた。反応液を室温にて16時間撹拌した後、エーテルで3回洗浄した。得られた水層がpH=3になるまで5M塩酸水溶液を加えた後、酢酸エチルで3回抽出した。有機溶媒を無水硫酸ナトリウムで乾燥させ、溶媒を減圧下留去した。得られた粗生成物をエーテルで洗浄することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-ブチルグリシン(化合物aa02、Fmoc-nBuGly-OH)(121 g)を得た。
LCMS(ESI)m/z = 354 (M+H)+
保持時間:0.87分(分析条件SQDFA05)


LCMS(ESI) m/z = 296 (M+Na)+
保持時間:1.39分(分析条件SMD method7)

得られたN-(tert-ブトキシカルボニル)-O-イソペンチル-L-セリン(Boc-Ser(iPen)-OH)(175 g)を1,4-ジオキサン(500 mL)に溶解し、濃塩酸(300 mL)を加えた後、室温で16時間撹拌した。反応液を減圧濃縮し、得られた粗生成物(122 g)を1,4-ジオキサン/水(1000 mL/1000 mL)に溶解し、炭酸カリウム(239 g、1.73 mol)およびN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(234 g、694.36 mmol)を加えて室温で16時間撹拌した。その後、反応液をヘキサンで3回洗浄した。洗浄後、塩酸水溶液(6M)を加えて水層のpHを2から3に調節し、酢酸エチルで2回抽出した。有機層を無水硫酸ナトリウムで乾燥し、ろ過後に減圧濃縮して溶媒を除去した。得られた粗生成物をエーテル、ヘキサン、酢酸エチルで洗浄し、さらに逆相カラムクロマトグラフィー(水/アセトニトリル=100/0→30/70)で精製してN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-イソペンチル-L-セリン(化合物aa04、Fmoc-Ser(iPen)-OH)(95 g、41%)を白色固体として得た。
LCMS(ESI) m/z = 398 (M+H)+
保持時間:2.29分(分析条件SMD method18)

上記のように得られた(S)-5-オキソ-4-フェネチルオキサゾリジン-3-カルボン酸(9H-フルオレン-9-イル)(50 g、118.51 mmol)のジクロロメタン溶液(700 mL)にトリエチルシラン(Et3SiH)、トリフルオロ酢酸(TFA)(700 mL)を加え、25℃で16時間撹拌した。反応液を減圧濃縮し、得られた残渣に炭酸カリウム水溶液を加え、石油エーテルで洗浄した。得られた水層を濃塩酸を用いてpH=3になるように調整し、酢酸エチルで抽出した。有機層を無水硫酸ナトリウムにて乾燥し、ろ過、減圧濃縮し、得られた残渣にメタノール、ヘキサンを加え再び減圧濃縮し、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-4-フェニルブタン酸(化合物aa05、Fmoc-MeHph-OH)(29.5 g、58%、2工程)を得た。
LCMS(ESI)m/z=416(M+H)+
保持時間:0.95分(分析条件SQDFA05)
N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((5-フルオロピリジン-3-イル)メチル)-L-セリン(化合物aa12、Fmoc-Ser(3-F-5-Me-Pyr)-OH)は下記のルートにて合成した。


LCMS(ESI)m/z=128(M+H)+
保持時間:0.28分(分析条件SQDFA05)

1H NMR(Varian400-MR、400MHz、d-DMSO)δ4.77(2H、s)、7.85-7.88(1H、m)、8.55-8.56(2H、m)

LCMS(ESI)m/z=455(M-H)-
保持時間:0.86分(分析条件SQDFA05)

LCMS(ESI)m/z=213(M-H)-
保持時間:0.24分(分析条件SQDAA05)
O-((5-フルオロピリジン-3-イル)メチル)-N-トリチル-L-セリン(化合物aa10、Trt-Ser(3-F-5-Me-Pyr)-OH)(98 mg、2.69 mmol)の1.4-ジオキサン溶液(819 μL)に5~10%の塩酸/メタノール溶液(2.45 mL)を室温にて加え、5時間撹拌した。反応液に1.4-ジオキサン(6.0 mL)を加え、得られた固体をろ過し、ジイソプロピルエーテルで洗浄、減圧下乾燥することでO-((5-フルオロピリジン-3-イル)メチル)-L-セリン(化合物aa11、H-Ser(3-F-5-Me-Pyr)-OH))(222.5 mg、86%)を塩酸塩として得た。
LCMS(ESI)m/z=213(M-H)-
保持時間:0.24分(分析条件SQDAA05)

LCMS(ESI)m/z=437(M+H)+
保持時間:0.86分(分析条件SQDAA05)
(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-クロロフェノキシ)プロパン酸(化合物aa16、Fmoc-Ser(Ph-3-Cl)-OH)は下記のルートにて合成した。


1H NMR(Varian400-MR、400MHz、d-DMSO)δ7.42-7.44(6H、m)、7.28-7.30(7H、m)、7.20-7.21(3H、m)、6.99-7.01(2H、m)、6.83-6.85(1H、m)、4.19-4.22(1H、m)、4.06-4.08(1H、m)、3.49-3.51(1H、m)、3.17(3H、s)

LCMS(ESI)m/z=216(M+H)+
保持時間:0.37分(分析条件SQDFA05)

LCMS(ESI)m/z=438(M+H)+
保持時間:0.91分(分析条件SQDFA05)

LCMS(ESI)m/z = 454 (M+H)+
保持時間:0.86分(分析条件SQDFA05)

LCMS(ESI) m/z = 388 (M+H)+
保持時間:1.13分(分析条件SMD method9)


LCMS(ESI) m/z = 372 (M+H)+
保持時間:1.31分(分析条件SMD method33)

LCMS(ESI)m/z = 456 (M+H)+
保持時間:0.81分(分析条件 SQDFA05)


LCMS(ESI)m/z = 487 (M+NH4)+
保持時間:0.68分(分析条件 SMD method11)

LCMS(ESI) m/z = 165 (M+H)+
保持時間:0.89分(分析条件SMD method12)


LCMS(ESI) m/z = 386 (M+H)+
保持時間:0.68分(分析条件SQDFA05)

LCMS(ESI) m/z = 468 (M-H)-
保持時間:0.86分、0.87分(分析条件SQDFA05)


LCMS(ESI) m/z = 501 (M+NH4)+
保持時間:0.82分(分析条件SMD method10)

LCMS(ESI) m/z = 163 (M-H)-
保持時間:7.21分(分析条件GC01)


LCMS(ESI) m/z = 386 (M+H)+
保持時間:2.08分(分析条件SMD method49)

LCMS(ESI) m/z = 468 (M-H)-
保持時間:0.85分、0.87分(分析条件SQDFA05)


LCMS(ESI) m/z = 501 (M+NH4)+
保持時間:0.82分(分析条件SMD method10)


LCMS(ESI) m/z = 416 (M+H)+
保持時間:0.93 分(分析条件 SQDFA05)

上で得られた、O-(2-(ベンジルオキシ)-2-メチルプロピル)-N-((ベンジルオキシ)カルボニル)-L-セリン(Cbz-Ser(tBuOBn)-OH)をメタノール(45 mL)に溶解し、Pd/C(456 mg)を加えて水素雰囲気化、室温で終夜撹拌した。セライトろ過によりPd/Cを除いた後、減圧濃縮によりメタノールを除去することでO-(2-ヒドロキシ-2-メチルプロピル)-L-セリン(H-Ser(tBuOH)-OH)(1.05 g)を定量的に得た。
得られたO-(2-ヒドロキシ-2-メチルプロピル)-L-セリン(H-Ser(tBuOH)-OH)(1.59 g、8.97 mL)および炭酸ナトリウム(2.85 g、26.9 mmol)を水(36 mL)および1,4-ジオキサン(15 mL)に溶解し、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(3.18 g、9.42 mmol)をゆっくり加えた。室温で30分撹拌後、反応液に水(40 mL)を加え、t-ブチルメチルエーテル(60 mL)で3回洗浄した。その後、水層に2Mの塩酸水溶液(26.9 mL、53.8 mmol)を加えて水層のpHを1とした後、t-ブチルメチルエーテル(60 mL)で3回抽出した。有機層を無水硫酸マグネシウムで乾燥し、ろ過後、t-ブチルメチルエーテルを減圧濃縮して得られた残渣を逆相カラムクロマトグラフィー(0.1%ギ酸水溶液/0.1ギ酸アセトニトリル溶液)で精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-ヒドロキシ-2-メチルプロピル)-L-セリン(化合物aa53、Fmoc-Ser(tBuOH)-OH)(2.62 g、73%)を得た。
LCMS(ESI) m/z = 400 (M+H)+
保持時間:0.71分(分析条件 SQDFA05)

LCMS(ESI) m/z = 484(M+H)+
保持時間:0.91分(分析条件SQDFA05)


得られた(S)-4-((2-ヒドロキシ-2-メチルプロポキシ)メチル)-5-オキソオキサゾリジン-3-カルボン酸(9H-フルオレン-9-イル)メチル(1 g、2.43 mmol)をトリフルオロ酢酸/ジクロロメタン(13 mL/13 mL)に溶解し、トリエチルシラン(Et3SiH)(832 mg、7.16 mmol)を滴下して加えた。室温で16時間撹拌後、反応液を減圧濃縮して得られた残渣を炭酸カリウム水溶液に溶解し、t-ブチルメチルエーテルで洗浄後、水層に塩酸水溶液(1M)を加えてpHを2に調節した。水層をt-ブチルメチルエーテルで2回抽出した後、有機層を水、飽和食塩水で各2回洗浄し、無水硫酸ナトリウムで乾燥させた。ろ過後、減圧濃縮して得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル=100/0→50/50)で精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-ヒドロキシ-2-メチルプロピル)-N-メチル-L-セリン(Fmoc-MeSer(tBuOH)-OH)(700 mg、70%)を得た。
得られたN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-ヒドロキシ-2-メチルプロピル)-N-メチル-L-セリン(Fmoc-MeSer(tBuOH)-OH)(6 g、14.51 mmol)とp-トルエンスルホン酸ピリジニウム(PPTS)(182 mg、0.73 mmol)をテトラヒドロフラン(48 mL)に溶解し、3,4-ジヒドロ-2H-ピラン(8.5 g、101.05 mmol)を窒素雰囲気化、室温で滴下して加えた。反応液を50度で5時間撹拌後、反応液に酢酸エチルを添加し、抽出した。有機層を無水硫酸ナトリウムで乾燥し、ろ過後、減圧濃縮してN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-O-(2-メチル-2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリンおよびテトラヒドロ-2H-ピラン-2-イルN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-O-(2-メチル-2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリナートを混合物として得た。
得られた混合物(50 g)をリン酸緩衝液(1M、pH=6.8、1000 mL)およびテトラヒドロフラン(1000 mL)に溶解させ、50度で4時間撹拌した。その後、反応液を酢酸エチルで抽出した。有機層を飽和食塩水で洗浄後、無水硫酸ナトリウムで乾燥し、ろ過後に減圧濃縮した。得られた残渣を逆相カラムクロマトグラフィー(0~40% 0.5%炭酸水素アンモニウム水溶液/アセトニトリル)で精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-O-(2-メチル-2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa55、Fmoc-MeSer(tBuOTHP)-OH)(20 g)を得た。
LCMS(ESI) m/z = 498 (M+H)+
保持時間:0.94分(分析条件SMD method20)


LCMS(ESI) m/z = 452 (M+Na)+
保持時間:1.46分(分析条件SMD method13)


LCMS(ESI) m/z = 498 (M+H)+
保持時間:0.72分(分析条件SMD method11)

LCMS(ESI) m/z = 448 (M+Na)+
保持時間:2.47分(分析条件SMD method14)

LCMS(ESI) m/z = 450 (M+Na)+
保持時間:2.12分(分析条件SMD method15)

得られた粗生成物である(S)-2-((tert-ブトキシカルボニル)アミノ)-6-ヒドロキシヘキサン酸メチル(Boc-Nle(6-OH)-OMe)(13 g)に4N塩酸/1.4-ジオキサン溶液(50 mL、200 mmol)を室温にて加え、6時間撹拌した。反応液をロータリーエバポレーターを用いて減圧濃縮することで、粗生成物として(S)-2-アミノ-6-ヒドロキシヘキサン酸メチル(H-Nle(6-OH)-OMe)の塩酸塩(9.2 g)を得た
得られた粗生成物である(S)-2-アミノ-6-ヒドロキシヘキサン酸メチル(H-Nle(6-OH)-OMe)の塩酸塩(9.2 g)のアセトニトリル溶液(100 mL)に1,3-ジオキソイソインドリン-2-カルボン酸エチル(9.74 g、44.4 mmol)、N,N-ジイソプロピルエチルアミン(DIPEA)(15.52 mL、89 mmol)を室温にて加え、終夜撹拌した。反応液をロータリーエバポレーターを用いて減圧濃縮することで、得られた残渣を順相カラムクロマトグラフィー(ジクロロメタン/メタノール)で精製することで(S)-2-(1,3-ジオキソイソインドリン-2-イル)-6-ヒドロキシヘキサン酸メチル(14.6 g)を得た。
上述の(S)-2-(1,3-ジオキソイソインドリン-2-イル)-6-ヒドロキシヘキサン酸メチル(7.08 g、24.3 mmol)のジクロロメタン溶液(100 mL)にデス-マーチンペルヨージナン(CAS#87413-09-0、11.34 g、26.7 mmol)を0度にて加え、室温にて3時間撹拌した。反応液をジクロロメタンで希釈し、飽和炭酸水素ナトリウム水溶液/水(1/1)の溶液、飽和チオ硫酸ナトリウム水溶液、飽和食塩水で洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、ろ過を行った後、減圧濃縮することで得られた残渣を順相カラムクロマトグラフィー(ヘキサン/酢酸エチル)で精製することで、(S)-2-(1,3-ジオキソイソインドリン-2-イル)-6-オキソヘキサン酸メチル(化合物aa65)(3.6 g、4工程51%)を得た。
LCMS(ESI)m/z = 290 (M+H)+
保持時間:0.70分(分析条件SQDAA05)

LCMS(ESI)m/z = 358 (M-H)-
保持時間:0.81分(分析条件SQDAA05)

上記の(2S)-2-アミノ-7,7,7-トリフルオロ-6-ヒドロキシヘプタン酸メチル(H-Hnl(7-F3-6-OH)-OMe)(2 g、8.73 mmol)に水(25 mL)、炭酸ナトリウム(2.93 g、34.9 mmol)、テトラヒドロフラン(50 mL)、炭酸N-スクシンイミジル9-フルオレニルメチル(Fmoc-OSu)(3.53 g、10.47 mmol)を室温にて加え、反応液を2時間撹拌した。テトラヒドロフランを除去するために反応液をロータリーエバポレーターで減圧濃縮し、酢酸エチル、1N塩酸水溶液を加え、酢酸エチルで2回抽出した。有機層を飽和食塩水で洗浄し、無水硫酸マグネシウムにて乾燥後、ろ過、減圧濃縮した。得られた残渣を逆相カラムクロマトグラフィー(0.1%ギ酸水溶液/0.1%ギ酸アセトニトリル溶液)、順相カラムクロマトグラフィー(ジクロロメタン/メタノール)で精製した後、さらに、逆相カラムクロマトグラフィー(10mM酢酸アンモニウム水溶液/メタノール)で精製することで得られたフラクションを集め、メタノールを減圧下留去し、酢酸エチルで2回抽出し、飽和硫酸水素カリウム溶液、飽和食塩水で洗浄した。得られた有機溶媒を減圧下留去しすることで、(2S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-7,7,7-トリフルオロ-6-ヒドロキシヘプタン酸メチル(化合物aa67、Fmoc-Hnl(7-F3-6-OH)-OMe)(1.4 g、2工程36%)を得た。
LCMS(ESI) m/z = 452 (M+H)+
保持時間:0.86分(分析条件SQDFA05)

LCMS(ESI) m/z = 438.6 (M+H)+
保持時間:0.76分(分析条件SQDFA05)

上記3つの反応液にそれぞれ飽和炭酸水素ナトリウム水溶液を加えた後、全ての反応液を合わせ減圧下、大部分の有機溶媒を留去し、残った溶液に酢酸エチル(300 mL)を加え抽出した。有機層を飽和食塩水溶液で洗浄した後、有機溶媒を減圧下留去した。得られた残渣を順相シリカゲルカラムクロマトグラフィー(ヘキサン/酢酸エチル)にて精製し、メチルN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3,3,3-トリフルオロ-2-ヒドロキシプロピル)-L-セリナート(化合物aa69、Fmoc-Ser(1-CF3-EtOH)-OMe)(6.57 g、53%、純度95%)および同化合物(3.1 g、25%、純度87%)を得た。
LCMS(ESI)m/z = 454 (M+H)+
保持時間:1.18分(分析条件SMD method45)

LCMS(ESI)m/z = 462 (M+Na)+
保持時間:2.24分(分析条件SMD method46)

ろ過した反応液にトリエチルシラン(2.0 mL、12.6 mmol)、三フッ化ホウ素ジエチルエーテル錯体(BF3・OEt2)(1.58 mL、12.6 mmol)を加え、室温で30分間撹拌した。反応液を飽和食塩水で希釈し、ジクロロメタンで抽出した。得られた有機層を減圧濃縮した後、得られた残渣を逆相カラムクロマトグラフィー(0.1%ギ酸水溶液/0.1%ギ酸アセトニトリル溶液)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-O-(3,3,3-トリフルオロ-2-ヒドロキシプロピル)-L-セリン(化合物aa71、Fmoc-MeSer(1-CF3-EtOH)-OH)(640 mg、34%)を得た。
LCMS(ESI) m/z = 452 (M-H)-
保持時間:2.33分(分析条件SMD method47)

LCMS(ESI)m/z=522(M-H)-
保持時間:1.00分(分析条件SQDAA05-2)

LCMS(ESI)m/z=324(M+H)+
保持時間:0.86分(分析条件SQDFA05)

LCMS(ESI) m/z = 439 (M+H)+
保持時間:1.05分(分析条件SMD method23)

LCMS(ESI) m/z = 383 (M+H)+
保持時間:1.75分(分析条件SMD method5)


LCMS(ESI) m/z = 441 (M+H)+
保持時間:1.08分(分析条件SMD method9)

窒素雰囲気下、上述の粗生成物である(S)-4-((2-(tert-ブチルアミノ)-2-オキソエトキシ)メチル)-5-オキソオキサゾリジン-3-カルボン酸(9H-フルオレン-9-イル)メチル(1.035 g)のジクロロメタン(DCM)溶液(2.86 mL)にトリエチルシラン(Et3SiH)(1.096 mL、6.86 mmol)と水(40 μL)を加え室温にて2分間撹拌した後、三フッ化ホウ素ジエチルエーテル錯体(BF3・OEt2)(0.580 mL、4.57 mmol)を加え、24時間室温にて撹拌した。反応液に飽和塩化アンモニウム水溶液を加え、室温で40分撹拌した後、ジクロロメタン(DCM)で2回抽出した。得られた有機層を無水硫酸ナトリウムで乾燥させ、減圧濃縮することで得られた残渣にアセトニトリルを加え、ヘキサンで洗浄した。アセトニトリルを減圧下留去することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-(tert-ブチルアミノ)-2-オキソエチル)-N-メチル-L-セリン(化合物aa79、Fmoc-MeSer(NtBu-Aca)-OH)(0.9449 g、2工程92%)を得た
LCMS(ESI)m/z = 455 (M+H)+
保持時間:0.79分(分析条件SQDFA05)


得られたO-(2-(メチルアミノ)-2-オキソエチル)-N-トリチル-L-セリン(Trt-Ser(NMe-Aca)-OH)(2.4 g、5.73 mmol)のジクロロメタン(DCM)溶液(2.87 mL)にトリフルオロ酢酸(TFA)(1.325 mL、17.20 mmol)を0度にて加え、30分間撹拌した。反応液に水(2.87 mL)を加え、得られた水層を逆相カラムクロマトグラフィー(水/アセトニトリル)で精製することで、O-(2-(メチルアミノ)-2-オキソエチル)-L-セリン(H-Ser(NtBu-Aca)-OH)(900 mg)を粗生成物として得た。これを精製することなく次工程に用いた。
得られた粗生成物であるO-(2-(メチルアミノ)-2-オキソエチル)-L-セリン(H-Ser(NtBu-Aca)-OH)(900 mg)に水(4.2 mL)、N-エチル-N-イソプロピルプロパン-2-アミン(DIPEA)(2.231 mL、12.77 mmol)の1,4-ジオキサン溶液(14.0 mL)を室温にて加えた後、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(1.723 g、5.11 mmol)を室温にて加え、30分間撹拌した。反応液を酢酸エチル(150 mL)、水(30 mL)、1N塩酸水溶液(3.0 mL)の混合液に加え、得られた有機層を減圧下留去した。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-(メチルアミノ)-2-オキソエチル)-L-セリン(化合物aa81、Fmoc-Ser(NMe-Aca)-OH)(1.8 g)を得た。
LCMS(ESI) m/z = 399 (M+H)+
保持時間:0.63分(分析条件SQDFA05)

LCMS(ESI) m/z = 334 (M+Na)+
保持時間:1.09分(分析条件SMD method13)


窒素雰囲気下、O-(3-ヒドロキシプロピル)-L-セリン(H-Ser(nPrOH)-OH)(5.5 g、33.707 mmol)、炭酸水素ナトリウム(8.6 g、102 mmol)の水/1,4-ジオキサン(50 mL/130 mL)溶液にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(11.94 g、35.396 mmol)を室温にて加え、4時間撹拌した。反応液に水を加え、t-ブチルメチルエーテルで2回洗浄した後、水層のpHが3になるまで6N塩酸水溶液を加え、t-ブチルメチルエーテルで抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過後、溶媒を減圧下留去し、さらに減圧ポンプで減圧下乾燥させた。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-ヒドロキシプロピル)-L-セリン(化合物aa84、Fmoc-Ser(nPrOH)-OH)(8 g、67%)を得た。
LCMS(ESI) m/z = 386 (M+H)+
保持時間:0.61分(分析条件SMD method50)

上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa85、Fmoc-Ser(nPrOTHP)-OH)とテトラヒドロ-2H-ピラン-2-イル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリナート(Fmoc-Ser(nPrOTHP)-OTHP)の混合物(1.5 g)にテトラヒドロフラン(8 mL)、1Mリン酸緩衝液(8 mL、pH 8.0)を加え、50度で3時間攪拌した。反応液を冷却し、t-ブチルメチルエーテルを加え、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った後、溶媒を減圧下留去した。得られた残渣をジエチルエーテル/ヘプタンで再結晶することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa85、Fmoc-Ser(nPrOTHP)-OH)をナトリウム塩(1 g)として得た。
上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa85、Fmoc-Ser(nPrOTHP)-OH)のナトリウム塩(5 g)にt-ブチルメチルエーテル(100 mL)、0.05Mリン酸水溶液(250 mL)を加え、室温で5分間撹拌した。反応液をt-ブチルメチルエーテルで2回抽出した後、飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った。溶媒を減圧下留去することで、粗生成物としてN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa85、Fmoc-Ser(nPrOTHP)-OH)(4.9 g)を得た。これをさらなる精製をすることなくペプチド合成に用いた。
LCMS(ESI)m/z = 468 (M―H)―
保持時間:0.84分(分析条件 SQDFA05)

LCMS(ESI) m/z = 326 (M+H)+
保持時間:1.12分(分析条件SMD method13)


窒素雰囲気下、O-(3-ヒドロキシ-2,2-ジメチルプロピル)-L-セリン(H-Ser(2-Me2-PrOH)-OH)(13 g、67.982 mmol)、炭酸水素ナトリウム(17.13 g、203.946 mmol)の水/1,4-ジオキサン(260 mL/104 mL)溶液にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(24.08 g、71.381 mmol)を室温にて加え、2時間撹拌した。反応液に水を加え、t-ブチルメチルエーテルで2回洗浄した後、水層に6N塩酸水溶液をpH3になるまで加え、t-ブチルメチルエーテルで抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過後、溶媒を減圧下留去し、さらに減圧ポンプで減圧下乾燥させた。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(3-ヒドロキシ-2,2-ジメチルプロピル)-L-セリン(化合物aa88、Fmoc-Ser(2-Me2-PrOH)-OH)(26.5 g、94%)を得た。
LCMS(ESI) m/z = 414 (M+H)+
保持時間:1.44分(分析条件SMD method51)

上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2,2-ジメチル-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa89、Fmoc-Ser(2-Me2-PrOTHP)-OH)とテトラヒドロ-2H-ピラン-2-イル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2,2-ジメチル-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリナート(Fmoc-Ser(2-Me2-PrOTHP)-OTHP)の混合物にテトラヒドロフラン(161 mL)、1Mリン酸緩衝液(161 mL、pH 8.0)を加え、50度で3時間攪拌した。反応液を冷却し、t-ブチルメチルエーテルを加え、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った後、溶媒を減圧下留去した。得られた残渣をジエチルエーテル/ヘプタンで再結晶することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2,2-ジメチル-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa89、Fmoc-Ser(2-Me2-PrOTHP)-OH)をナトリウム塩(17.1 g)として得た。
上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2,2-ジメチル-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa89、Fmoc-Ser(2-Me2-PrOTHP)-OH)のナトリウム塩(8 g)にt-ブチルメチルエーテル(120 mL)、0.05Mリン酸水溶液(350 mL)を加え、室温で5分間撹拌した。反応液をt-ブチルメチルエーテルで抽出し、飽和食塩水で2回洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。溶媒を減圧下留去することで、粗生成物としてN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2,2-ジメチル-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)プロピル)-L-セリン(化合物aa89、Fmoc-Ser(2-Me2-PrOTHP)-OH)(7 g)を得た。これをさらなる精製をすることなくペプチド合成に用いた。
LCMS(ESI)m/z = 496 (M―H)―
保持時間:0.97分(分析条件 SQDFA05)

LCMS(ESI) m/z = 179 (M+H)+
保持時間:1.26分(分析条件SMD method13)

LCMS(ESI) m/z = 181 (M+H)+
保持時間:1.16分(分析条件SMD method13)

LCMS(ESI) m/z = 416 (M+H)+
保持時間:1.39分(分析条件SMD method13)


窒素雰囲気下、上述のようにして得たO-((S)-3-ヒドロキシブチル)-L-セリン(H-Ser(S-2-BuOH)-OH)(5.85 g、33.03 mmol)、炭酸水素ナトリウム(8.30 g、98.81 mmol)の水/1,4-ジオキサン(135 mL/55 mL)溶液にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(11.7 g、34.6815 mmol)を室温にて加え、2時間撹拌した。反応液に水を加え、t-ブチルメチルエーテルで2回洗浄した後、水層のpHが3になるまで6N塩酸水溶液を加え、t-ブチルメチルエーテルで抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過後、溶媒を減圧下留去し、さらに減圧ポンプで減圧下乾燥させた。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((S)-3-ヒドロキシブチル)-L-セリン(化合物aa94、Fmoc-Ser(S-2-BuOH)-OH)(8.1 g、61%)を得た。
LCMS(ESI) m/z = 400 (M+H)+
保持時間:1.21分(分析条件SMD method13)

上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3S)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa95、Fmoc-Ser(S-2-BuOTHP)-OH)とテトラヒドロ-2H-ピラン-2-イル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3S)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリナート(Fmoc-Ser(S-2-BuOTHP)-OTHP)の混合物にテトラヒドロフラン(90 mL)、1Mリン酸緩衝液(90 mL、pH 8.0)を加え、50度で3時間攪拌した。反応液を冷却し、t-ブチルメチルエーテルを加え、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った後、溶媒を減圧下留去した。得られた残渣をジエチルエーテル/ヘプタンで再結晶することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3S)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa95、Fmoc-Ser(S-2-BuOTHP)-OH)をナトリウム塩(7.4 g)として得た。
上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3S)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa95、Fmoc-Ser(S-2-BuOTHP)-OH)のナトリウム塩(7.4 g)にt-ブチルメチルエーテル(130 mL)、0.05Mリン酸水溶液(360 mL)を加え、室温で5分間撹拌した。反応液をt-ブチルメチルエーテルで抽出し、飽和食塩水で2回洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。溶媒を減圧下留去することで、粗生成物としてN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3S)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa95、Fmoc-Ser(S-2-BuOTHP)-OH)(6.9 g)を得た。これをさらなる精製をすることなくペプチド合成に用いた。
LCMS(ESI)m/z = 482 (M―H)―
保持時間:0.87分、0.88分(分析条件 SQDFA05)

LCMS(ESI) m/z = 179 (M+H)+
保持時間:1.29分(分析条件SMD method13)

LCMS(ESI) m/z = 181 (M+H)+
保持時間:1.11分(分析条件SMD method13)

LCMS(ESI) m/z = 416 (M+H)+
保持時間:1.42分(分析条件SMD method13)


窒素雰囲気下、上述のようにして得たO-((R)-3-ヒドロキシブチル)-L-セリン(H-Ser(R-2-BuOH)-OH)(5.3 g、29.91 mmol)、炭酸水素ナトリウム(7.54 g、89.73 mmol)の水/1,4-ジオキサン(134 mL/57 mL)溶液にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(10.58 g、31.405 mmol)を室温にて加え、2時間撹拌した。反応液に水を加え、t-ブチルメチルエーテルで2回洗浄した後、水層のpHが3になるまで6N塩酸水溶液を加え、t-ブチルメチルエーテルで抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過後、溶媒を減圧下留去し、さらに減圧ポンプで減圧下乾燥させた。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製し、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((R)-3-ヒドロキシブチル)-L-セリン(化合物aa100、Fmoc-Ser(R-2-BuOH)-OH)(7.4 g、62%)を得た。
LCMS(ESI) m/z = 400 (M+H)+
保持時間:1.20分(分析条件SMD method13)

上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3R)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa101、Fmoc-Ser(R-2-BuOTHP)-OH)とテトラヒドロ-2H-ピラン-2-イル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3R)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリナート(Fmoc-Ser(R-2-BuOTHP)-OTHP)の混合物にテトラヒドロフラン(74 mL)、1Mリン酸緩衝液(74 mL、pH 8.0)を加え、50度で3時間攪拌した。反応液を冷却し、t-ブチルメチルエーテルを加え、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った後、溶媒を減圧下留去した。得られた残渣をジエチルエーテル/ヘプタンで再結晶することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3R)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa101、Fmoc-Ser(R-2-BuOTHP)-OH)をナトリウム塩(10.2 g)として得た。
上述のようにして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3R)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa101、Fmoc-Ser(R-2-BuOTHP)-OH)のナトリウム塩(10 g)にt-ブチルメチルエーテル(160 mL)、0.05Mリン酸水溶液(440 mL)を加え、室温で5分間撹拌した。反応液をt-ブチルメチルエーテルで抽出し、飽和食塩水で2回洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。溶媒を減圧下留去することで、粗生成物としてN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((3R)-3-((テトラヒドロ-2H-ピラン-2-イル)オキシ)ブチル)-L-セリン(化合物aa101、Fmoc-Ser(R-2-BuOTHP)-OH)(8.8 g)を得た。これをさらなる精製をすることなくペプチド合成に用いた。
LCMS(ESI)m/z = 482 (M―H)―
保持時間:0.86分、0.88分(分析条件 SQDFA05)

LCMS(ESI) m/z = 434 (M+H)+
保持時間:0.73分(分析条件SQD2FA05)


塩化カルシウム(5.98 g、53.8 mmol)に水(14.96 mL)、水酸化リチウムの1水和物(0.603 g、14.36 mmol)を加え、室温で5分間撹拌した。反応液にイソプロパノール(59.8 mL)と(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(4-((2-クロロフェニル)ジフェニルメトキシ)-3-メトキシフェニル)プロパン酸メチル(Fmoc-Phe(4-OClt-3-OMe)-OMe)(2.6 g、3.59 mmol)のテトラヒドロフラン(THF)溶液(14.96 mL)を加え、4時間撹拌した後、0.05Mリン酸水溶液とt-ブチルメチルエーテルを加えた。水層をt-ブチルメチルエーテルで3回抽出し、得られた有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥させ、ろ過を行った後、減圧下溶媒留去した。粗生成物として得られた(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(4-((2-クロロフェニル)ジフェニルメトキシ)-3-メトキシフェニル)プロパン酸(化合物aa104、Fmoc-Phe(4-OClt-3-OMe)-OH)(2.8 g)はさらなる精製を行わずにペプチド合成に用いた。
LCMS(ESI) m/z = 708 (M―H)―
保持時間:0.74分(分析条件SQD2AA50)
N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa109、Fmoc-Ser(3-Me-5-Oxo-Odz)-OH)は下記のスキームに従って合成した。

上記のようにして得たN-(tert-ブトキシカルボニル)-O-(シアノメチル)-L-セリン(化合物aa105、Boc-Ser(CH2CN)-OH)(38.5 g、157.63 mmol)のエタノール(EtOH)溶液(400 mL)に室温にてヒドロキシルアミン塩酸塩(22.4 g、315.26 mmol)およびトリエチルアミン(58.8 mL、425.60 mmol)を加え、85度で2時間撹拌した。減圧下溶媒を留去し、粗生成物であるN-(tert-ブトキシカルボニル)-O-(2-(ヒドロキシアミノ)-2-イミノエチル)-L-セリン(化合物aa106)(107 g)を得た。
窒素雰囲気下、得られた粗生成物であるN-(tert-ブトキシカルボニル)-O-(2-(ヒドロキシアミノ)-2-イミノエチル)-L-セリン(化合物aa106)(12 g)の1、4-ジオキサン溶液(120 mL)に室温にてカルボニルジイミダゾール(CDI)(3.96 g、24.42 mmol)、1,8-ジアザビシクロ[5.4.0]-7-ウンデセン(DBU)(4.92 g、32.32 mmol)を加え、110度で2時間撹拌した。濃塩酸を用い、反応液をpH=2に調節した後、ジクロロメタン(DCM)で2回抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過を行ったを行った後、ろ液を減圧下濃縮することで、粗生成物としてN-(tert-ブトキシカルボニル)-O-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa107、Boc-Ser(3-Me-5-Oxo-Odz)-OH)(1.2 g)を得た。
窒素雰囲気下、得られた粗生成物であるN-(tert-ブトキシカルボニル)-O-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa107、Boc-Ser(3-Me-5-Oxo-Odz)-OH)(258 mg)の1、4-ジオキサン溶液(10 mL)に室温にて4N塩酸/1,4-ジオキサン溶液(15 mL)を加え、16時間撹拌した。減圧下溶媒を留去し、粗生成物であるO-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa108、H-Ser(3-Me-5-Oxo-Odz)-OH)(160 mg)を得た。
窒素雰囲気下、上記のように得た粗生成物であるO-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa108、H-Ser(3-Me-5-Oxo-Odz)-OH)(22.4 g)と炭酸カリウム(18.8 g、135.04 mmol)の1.4-ジオキサン/水(165 mL/110 mL)溶液にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(23 g、68.2 mmol)を室温にて加え、3時間撹拌した。反応液をt-ブチルメチルエーテル/ヘキサン(1/3)で洗浄し、得られた水層を濃塩酸を用いてpH=2になるまで調節した後、ジクロロメタン(DCM)で2回抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-((5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)メチル)-L-セリン(化合物aa109、Fmoc-Ser(3-Me-5-Oxo-Odz)-OH)(5.69 g)を得た。
LCMS(ESI)m/z = 426 (M+H)+
保持時間:0.72分(分析条件SQD2FA05)

LCMS(ESI)m/z=502(M+Na)+
保持時間:0.87分(分析条件SQDFA05)
約50%の水が含まれる1-メチル-3-ニトロ-1-ニトロソグアニジン(5.0 g、東京化成あるいは富士フィルム和光純薬より購入)にジエチルエーテル(17.0 mL)を加え0度にて50%水酸化カリウム水溶液(9.5 mL)を滴下しながら加えた。反応液を1時間撹拌した後、黄色のジエチルエーテル層を先端をなめしたパスツールを用い分離し、得られた溶液をそのまま上記の反応に使用した。

LCMS(ESI)m/z=468(M―H)―
保持時間:0.78分(分析条件SQDFA05)

窒素雰囲気下、論文記載の方法(Beilstein Journal of Organic Chemistry, 2017, 13, 2428-2441.)で合成した2-(トリチルオキシ)エタン-1-オール(CAS. No. 18325-45-6)(8.8 g、28.9 mmol)と市販の4-メチルベンゼンスルホン酸 (R)-オキシラン-2-イルメチル(6.0 g、26.3 mmol)のジメチルホルムアミド(DMF)溶液(100 mL)に0度にて水素化ナトリウム(1.16 g、28.9 mmol、60%オイルディスパージョン)を加え、室温にて12時間撹拌した。反応液に水を加えた後、酢酸エチルで2回抽出し、得られた有機層を減圧下濃縮し、得られた残渣を順相カラムクロマトグラフィー(ヘキサン/酢酸エチル=5/1)にて精製することで、(R)-2-((2-(トリチルオキシ)エトキシ)メチル)オキシラン(化合物aa112)(7.32 g、77%)を得た。
LCMS(ESI)m/z = 383 (M+Na)+
保持時間:1.40分(分析条件SMD method45)
LCMS(ESI)m/z = 500 (M+Na)+
保持時間:1.40分(分析条件SMD method45)
LCMS(ESI)m/z = 578 (M+Na)+
保持時間:1.47分(分析条件SMD method45)
LCMS(ESI)m/z = 509 (M+Na)+
保持時間:1.49分(分析条件SMD method45)
LCMS(ESI)m/z = 386 (M+H)+
保持時間:1.78分(分析条件SMD method47)
LCMS(ESI)m/z = 492 (M+Na)+
保持時間:0.95分(分析条件SQDAA05-2)

窒素雰囲気下、市販の(S)-4,4-ジフルオロピロリジン-2-カルボン酸メチル塩酸塩(CAS. No. 156046-05-8)(6.9 g、34.2 mmol)のジクロロメタン溶液(68.5 mL)に室温にてトリエチルアミン(11.9 mL、86 mmol)を加えた。反応液に二炭酸ジ-tert-ブチル(Boc2O)(11.2 g、51.3 mmol)を室温にて加え、1時間撹拌した。反応液に水を加えた後、酢酸エチルで抽出した。得られた有機層を減圧下濃縮し、得られた残渣を順相カラムクロマトグラフィー(n-ヘキサン/酢酸エチル)にて精製することで、(S)-4,4-ジフルオロピロリジン-1,2-ジカルボン酸 2-メチル 1-(tert-ブチル)(化合物aa118)(6.85 g、75%)を得た。
LCMS(ESI)m/z = 567 (M+Na)+
保持時間:1.32分(分析条件SMD method45)
LCMS(ESI)m/z = 529 (M+H)+
保持時間:2.75分(分析条件SMD method47)

LCMS(ESI)m/z = 250 (M+H)+
保持時間:0.91分(分析条件SMD method45)
LCMS(ESI)m/z = 222 (M+H)+
保持時間:0.76分(分析条件SMD method45)
LCMS(ESI)m/z = 545(M+H)+
保持時間:3.02分(分析条件SMD method47)
LCMS(ESI)m/z = 531 (M+H)+
保持時間:2.69分(分析条件SMD method47)


LCMS(ESI)m/z = 574 (M+NH4)+
保持時間:1.00分(分析条件SQD2FA05)

LCMS(ESI)m/z = 292 (M+H)+
保持時間:0.78分(分析条件SMD method50)

窒素雰囲気下、上述のように得た(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-メトキシ-4-(メチルカルバモイル)フェニル)プロパン酸 tert-ブチル(Fmoc-Phe(3-OMe-4-CONMe)-OtBu)(11 g、20.8 mmol)とクロロトリメチルシラン(TMSCl)(7.96 mL、62.3 mmol)の2,2,2-トリフルオロエタノール(TFE)溶液(104 mL)を室温にて1時間撹拌した。反応液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-メトキシ-4-(メチルカルバモイル)フェニル)プロパン酸(化合物aa127、Fmoc-Phr(3-OMe-4-CONMe)-OH)(5.0 g、51%)を得た。
LCMS(ESI)m/z = 475 (M+H)+
保持時間:0.68分(分析条件SQDFA05)

LCMS(ESI)m/z = 292 (M+H)+
保持時間:0.79分(分析条件SMD method50)

窒素雰囲気下、上述のように得た(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(4-メトキシ-3-(メチルカルバモイル)フェニル)プロパン酸 tert-ブチル(Fmoc-Phe(4-OMe-3-CONMe)-OtBu)(10.2 g、19.22 mmol)とクロロトリメチルシラン(TMSCl)(7.26 mL、56.8 mmol)の2,2,2-トリフルオロエタノール(TFE)溶液(96 mL)を室温にて1時間撹拌した。反応液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(4-メトキシ-3-(メチルカルバモイル)フェニル)プロパン酸(化合物aa128、Fmoc-Phe(4-OMe-3-CONMe)-OH)(6.12 g、36%)を得た。
LCMS(ESI)m/z = 475 (M+H)+
保持時間:0.71分(分析条件SQDFA05)

窒素雰囲気下、臭化ニッケル(II)の3水和物(NiBr2・3H2O)(2.93 g、10.78 mmol)と4,4'-ジ-tert-ブチル-2,2'-ビピリジル(dtbbpy)(2.89 g、10.78 mmol)のジメチルアセトアミド(DMA)溶液(100 mL)を50℃で3時間撹拌した後、室温に冷却することでニッケル触媒溶液を得た。窒素雰囲気下、亜鉛粉末(11.75 g、179.672 mmol)、(((9H-フルオレン-9-イル)メトキシ)カルボニル)-L-アスパラギン酸 4-(1,3-ジオキソイソインドリン-2-イル) 1-(tert-ブチル)(化合物aa130)(20.0 g、35.934 mmol)、4-ブロモ-2-メトキシ-N-(メチルスルホニル)ベンズアミド(33.22 g、107.803 mmol)のジメチルアセトアミド(DMA)溶液(100 mL)を室温撹拌した後、上述のニッケル触媒溶液を滴下しながら室温にて加え、16時間撹拌した。反応液に10%EDTA・2Na(エチレンジアミン四酢酸の2ナトリウム塩)水溶液を加え、酢酸エチルで3回抽出した。得られた有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥させた後、ろ過、減圧濃縮を行うことで得られた残渣を順相カラムクロマトグラフィー(石油エーテル/酢酸エチル)にて精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-メトキシ-4-((メチルスルホニル)カルバモイル)フェニル)プロパン酸 tert-ブチル(Fmoc-Phe(3-OMe-4-CONHMs)-OtBu)(5.05g、19%)を得た。
窒素雰囲気下、上述のように得た(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-メトキシ-4-((メチルスルホニル)カルバモイル)フェニル)プロパン酸 tert-ブチル(Fmoc-Phe(3-OMe-4-CONHMs)-OtBu)(10.1 g、16.984 mmol)とクロロトリメチルシラン(TMSCl)(5.54 g、50.952 mmol)の2,2,2-トリフルオロエタノール(TFE)溶液(100 mL)を室温にて1時間撹拌した。反応液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-(3-メトキシ-4-((メチルスルホニル)カルバモイル)フェニル)プロパン酸(化合物aa129、Fmoc-Phe(3-OMe-4-CONHMs)-OH)(5.06 g、55%)を得た。
LCMS(ESI)m/z = 539 (M+H)+
保持時間:0.72分(分析条件SQDFA05)

水素雰囲気下、得られた粗生成物である(2S,4R)-1-((ベンジルオキシ)カルボニル)-4-(2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)エトキシ)ピロリジン-2-カルボン酸(Cbz-Hyp(2-EtOTHP)-OH)(10.1 g)とパラジウム炭素(Pd/C)(3.0 g)のメタノール溶液(100 mL)を室温にて2時間撹拌した。反応液をろ過した後、得られた固体をメタノール(100 mL)で洗浄した。ろ液を減圧下濃縮することで得られた残渣をエーテルで洗浄することで(2S,4R)-4-(2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)エトキシ)ピロリジン-2-カルボン酸(H-Hyp(2-EtOTHP)-OH)(6.1 g)を粗生成物として得た。
得られた粗生成物である(2S,4R)-4-(2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)エトキシ)ピロリジン-2-カルボン酸(H-Hyp(2-EtOTHP)-OH)(6.1 g)と炭酸水素ナトリウム(5.9 g、70.5 mmol)の1、4-ジオキサン/水=1/1溶液(130 mL)にN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(7.9 g、23.5 mmol)を加え、室温にて2時間撹拌した。反応液にt-ブチルメチルエーテル(TBME)/ヘキサン=1/3溶液(100 mL)を加え、洗浄する操作を3回行った後、得られた水層に1N塩酸水溶液をpH=6になるまで加え、t-ブチルメチルエーテル(TBME)(100 mL)で抽出した。得られた有機層を飽和食塩水で洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。得られたろ液を減圧下濃縮し、残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、(2S,4R)-1-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-4-(2-((テトラヒドロ-2H-ピラン-2-イル)オキシ)エトキシ)ピロリジン-2-カルボン酸(化合物aa133、Fmoc-Hyp(2-EtOTHP)-OH)(1.88 g、3工程30%)で得た。
LCMS(ESI)m/z = 482 (M+H)+
保持時間:0.80分(分析条件SQDFA05)

文献記載の方法(Journal of Medicinal Chemistry (1991), 34(4), 1297-1301)により調整した3-((2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸メチル(31.0 g、142.683 mmol)とN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(48.13 g、142.683 mmol)の1,4-ジオキサン溶液(200 mL)に炭酸ナトリウム(30.25 g、285.366 mmol)の水溶液(200 mL)を室温にて加え16時間撹拌した。反応液にアセトニトリルを加え、ろ過をした後、ろ液を減圧下濃縮した。得られた残渣に酢酸エチルを加え、飽和食塩水で洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。
得られたろ液を減圧下濃縮することで粗生成物として3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸メチル(62 g、36%)を得た。
塩化カルシウム(84.54 g、761.761 mmol)と水酸化リチウムの1水和物(8.52 g、203.136 mmol)に水(250 mL)とイソプロパノール(1000 mL)を加えた後、上述の粗生成物である3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸メチル(62 g、50.784 mmol)のテトラヒドロフラン溶液(250 mL)を0℃にて加え、16時間撹拌した。反応液を減圧下濃縮した後、得られた残渣に水を加え、t-ブチルメチルエーテル(TBME)で洗浄した。得られた水層に1N塩酸水溶液を溶液のpHが酸性になるまで加えた後、酢酸エチルで抽出した。得られた有機層を水、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った。得られたろ液を減圧下濃縮することで粗生成物として3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸(14 g、58%)を得た。
窒素雰囲気下、上述の粗生成物である3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸(14 g、32.904 mmol)、N-ヒドロキシフタルイミド(5.90 g、36.194 mmol)、N,N'-ジシクロヘキシルカルボジイミド(DCC)(7.47 g、36.194 mmol)のテトラヒドロフラン溶液(140 mL)を室温にて16時間撹拌した。反応液をろ過した後、減圧下濃縮することで得られた残渣に酢酸エチルを加え、飽和食塩水で洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。得られたろ液を減圧下濃縮することで粗生成物として3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸 1,3-ジオキソイソインドリン-2-イル(14 g、75%)を得た。
窒素雰囲気下、臭化ニッケル(II)の3水和物(NiBr2・3H2O)(5.73 g、21.031mmol)と4,4'-ジ-tert-ブチル-2,2'-ビピリジル(dtbbpy)(5.64 g、21.031 mmol)のジメチルアセトアミド(DMA)溶液(200 mL)を50℃で3時間撹拌した後、室温に冷却することでニッケル触媒溶液を得た。窒素雰囲気下、亜鉛粉末(22.93 g、350.509 mmol)、上述の様にして得た粗生成物である3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(2-(tert-ブトキシ)-2-オキソエチル)アミノ)プロパン酸 1,3-ジオキソイソインドリン-2-イル(40.0 g、70.102 mmol)、実施例に記載の通り合成した4-ヨード-2-メトキシ-N-メチルベンズアミド(化合物aa131)(40.81 g、140.204 mmol)のジメチルアセトアミド(DMA)溶液(200 mL)を室温で10分間撹拌した後、上述のニッケル触媒溶液を滴下しながら室温にて加え、16時間撹拌した。反応液に10%EDTA・2Na(エチレンジアミン四酢酸の2ナトリウム塩)水溶液を加え、ろ過した後、酢酸エチルで3回抽出した。有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った後、得られたろ液を減圧下濃縮し、残渣を順相カラムクロマトグラフィーにて精製することで、tert-ブチル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-(3-メトキシ-4-(メチルカルバモイル)フェネチル)グリシナート(14 g、37%)を得た。
上述のtert-ブチル N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-(3-メトキシ-4-(メチルカルバモイル)フェネチル)グリシナート(14.0 g、25.705 mmol)の4N塩酸/1,4-ジオキサン溶液(65 mL)を室温にて48時間撹拌した。反応液を減圧下濃縮した後、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-(3-メトキシ-4-(メチルカルバモイル)フェネチル)グリシン(化合物aa134、Fmoc-(Ph(3-OMe-4-CONMe)Et)Gly-OH)(6 g、47%)を得た。
LCMS(ESI)m/z = 489 (M+H)+
保持時間:0.70分(分析条件SQDFA05)

窒素雰囲気下、市販の(S)-4-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-5-(tert-ブトキシ)-5-オキソペンタン酸(Fmoc-Glu-OtBu)(175 g、411.299 mmol)、N-ヒドロキシフタルイミド(67.1 g、411.299 mmol)、1-エチル-3-(3-ジメチルアミノプロピル)カルボジイミド塩酸塩(EDCI・HCl)(78.85 g、411.299 mmol)、N,N-ジメチル-4-アミノピリジン(DMAP)(2.51 g、20.565 mmol)のジメチルホルムアミド溶液(2000 mL)を室温にて16時間撹拌した。反応液に1 M塩酸水溶液を加えた後、t-ブチルメチルエーテル(TBME)で抽出した。得られた有機層を水、50%飽和炭酸水素ナトリウム水溶液、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った後、得られたろ液を減圧下濃縮し、残渣を順相カラムクロマトグラフィー(石油エーテル/酢酸エチル)にて精製することで、(((9H-フルオレン-9-イル)メトキシ)カルボニル)-L-グルタミン酸 5-(1,3-ジオキソイソインドリン-2-イル) 1-(tert-ブチル)(105 g、45%)を得た。
窒素雰囲気下、臭化ニッケル(II)の3水和物(NiBr2・3H2O)(141 mg、0.526 mmol)と4,4'-ジ-tert-ブチル-2,2'-ビピリジル(dtbbpy)(140 mg、0.526 mmol)のジメチルアセトアミド(DMA)溶液(5 mL)を50℃で3時間撹拌した後、室温に冷却することでニッケル触媒溶液を得た。窒素雰囲気下、亜鉛粉末(617 mg、8.75 mmol)、上述の様にして得た(((9H-フルオレン-9-イル)メトキシ)カルボニル)-L-グルタミン酸 5-(1,3-ジオキソイソインドリン-2-イル) 1-(tert-ブチル)(1.0 g、1.75 mmol)、実施例に記載の通り合成した4-ヨード-2-メトキシ-N-メチルベンズアミド(化合物aa131)(1.19 g、3.50 mmol)のジメチルアセトアミド(DMA)溶液(5 mL)を室温で1時間撹拌した後、上述のニッケル触媒溶液を滴下しながら室温にて加え、16時間撹拌した。反応液に10%EDTA・2Na(エチレンジアミン四酢酸の2ナトリウム塩)水溶液を加え、ろ過した後、酢酸エチルで3回抽出した。有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った後、得られたろ液を減圧下濃縮し、残渣を順相カラムクロマトグラフィー(石油エーテル/酢酸エチル)にて精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-4-(3-メトキシ-4-(メチルカルバモイル)フェニル)ブタン酸 tert-ブチル(550 mg、57%)を得た。
上述の様にして得た(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-4-(3-メトキシ-4-(メチルカルバモイル)フェニル)ブタン酸 tert-ブチル(3.00 g、5.508 mmol)のトリフルオロエタノール(TFE)溶液(30 mL)に室温にてクロロトリメチルシラン(TMS-Cl)(1.80 g、16.568 mmol)を加え、3時間撹拌した。反応液を減圧下濃縮した後、得られた残渣を酢酸エチルで精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-4-(3-メトキシ-4-(メチルカルバモイル)フェニル)ブタン酸(化合物aa135、Fmoc-Hph(3-OMe-4-CONMe)-OH)(1.9 g、70%)を得た。
LCMS(ESI)m/z = 489 (M+H)+
保持時間:0.71分(分析条件SQDFA05)

LCMS(ESI)m/z = 487 (M―H)―
保持時間:0.83分(分析条件SQDFA05)

LCMS(ESI)m/z = 503 (M+H)+
保持時間:0.73分(分析条件SQDFA05)

窒素雰囲気下、上述の粗生成物として得られた(S)-4-(3-(アリルオキシ)-3-オキソプロピル)-5-オキソオキサゾリジン-3-カルボン酸 (9H-フルオレン-9-イル)メチル(110 g)、トリエチルシラン(Et3SiH)(60.6 g、522.565 mmol)、水(4.7 mL、264.282 mmol)のジクロロメタン溶液(1100 mL)に0℃にて三フッ化ホウ素ジエチルエーテル錯体(BF3・OEt2)(74.2 g、522.565 mmol)を加え、16時間撹拌した。反応液に塩化アンモニウム水溶液を加え、酢酸エチルで2回抽出した後、得られた有機層を減圧下濃縮した。残渣に炭酸水素ナトリウム水溶液を加え溶解させた後、ヘキサン/酢酸エチル(5/1)で3回洗浄した。水層に6N塩酸水溶液をpH=2になるまで加え、酢酸エチルで2回抽出した。有機層を水、飽和食塩水で洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った後、ろ液を減圧下濃縮した。得られた残渣を1%トリフルオロ酢酸(TFA)のアセトニトリル/水(1/5)溶液を用いて再結晶することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-5-(アリルオキシ)-5-オキソペンタン酸(Fmoc-MeGlu(OAll)-OH)(95 g、2工程86%)を得た。
窒素雰囲気下、上述の様に得た(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-5-(アリルオキシ)-5-オキソペンタン酸(Fmoc-MeGlu(OAll)-OH)(100 g、236.406 mmol)と三フッ化ホウ素ジエチルエーテル錯体(BF3・OEt2)(0.16 g、2.364 mmol)のジクロロメタン溶液(260 mL)に0℃にて2,2,2-トリクロロアセトイミド酸 tert-ブチル(51 g、472.813 mmol)のシクロヘキサン溶液を加えた後、1時間撹拌した。反応液にピリジンを加え、ろ過を行った後、ろ液を炭酸水素ナトリウム水溶液、飽和食塩水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮することで得られた残渣をアセトニトリル/水(1/5)溶液を用いて再結晶することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-L-グルタミン酸 1-(tert-ブチル) 5-アリル(102 g)を定量的に得た。
窒素雰囲気下、上述の様にして得たN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチル-L-グルタミン酸 1-(tert-ブチル) 5-アリル(290 g、605.427 mmol)とテトラキス(トリフェニルホスフィン)パラジウム(0)(Pd(PPh3)4)(7.0 g、6.054 mmol)のジクロロメタン溶液(1450 mL)にフェニルシラン(PhSiH3)(45.77 g、423.799 mmol)を滴下しながら室温にて加え、撹拌した。原料の消失をLC-MSにて確認した後、反応液をろ過し、得られたろ液を減圧下濃縮した。残渣に炭酸ナトリウム水溶液を加え溶解させた後、ヘキサン/t-ブチルメチルエーテル(1/3)溶液で3回洗浄した。水層にリン酸をpH=2になるまで加えた後、t-ブチルメチルエーテルで2回抽出した。有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った後、得られたろ液を減圧下濃縮することで、(S)-4-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-5-(tert-ブトキシ)-5-オキソペンタン酸 (化合物aa138、Fmoc-MeGlu-OtBu)(247 g、93%)を得た。
LCMS(ESI)m/z = 462 (M+Na)+
保持時間:1.380分(分析条件SMD method52)


LCMS(ESI)m/z = 545 (M+H)+
保持時間:0.79分(分析条件SQDFA05)


LCMS(ESI)m/z = 561 (M+H)+
保持時間:0.77分(分析条件SQDFA05)

窒素雰囲気下、市販の(S)-モルホリン-3-カルボン酸(CAS. No. 741288-31-3)(14.0 g、107 mmol)の水溶液(100 mL)に室温にて炭酸水素ナトリウム(20.26 g、241.172 mmol)を加えた。反応液に室温でクロロギ酸アリル(Alloc-Cl)(17.44 g、120.53 mmol)の1,4-ジオキサン溶液(100 mL)を滴下しながら加え、2時間撹拌した。反応液に水を加えた後、酢酸エチルで抽出し、得られた有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を順相カラムクロマトグラフィー(石油エーテル/酢酸エチル)にて精製することで、(S)-モルホリン-3,4-ジカルボン酸 3-メチル 4-アリル(19 g、86%)を得た。
上記の様にして得た(S)-モルホリン-3,4-ジカルボン酸 3-メチル 4-アリル(19.5 g、85.067 mmol)のテトラヒドロフラン(THF)溶液(283 mL)に0度にてリチウムホウ水素化物(5.56 g、255.233)/テトラヒドロフラン(THF)溶液を滴下しながら加え、室温で2時間撹拌した。反応液に0度にて水と2N塩酸水溶液を加えた後、酢酸エチルで抽出し、得られた有機層を飽和食塩水で洗浄、無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を順相カラムクロマトグラフィー(石油エーテル/酢酸エチル)にて精製することで、(R)-3-(ヒドロキシメチル)モルホリン-4-カルボン酸アリル(10.1 g、59%)を得た。
窒素雰囲気下、上述の(R)-3-(ヒドロキシメチル)モルホリン-4-カルボン酸アリル(7.0 g、34.8 mmol)と特許記載の方法(WO2015/179823)で合成した(S)-アジリジン-1,2-ジカルボン酸 2-メチル 1-((9H-フルオレン-9-イル)メチル)(7.50 g、23.2 mmol)のジクロロメタン溶液(47 mL)に0度にて三フッ化ホウ素-エチルエーテルコンプレックス(1.48 g、10.45 mmol)のジクロロメタン溶液を滴下しながら加え、2時間撹拌した。反応液に水を加えた後、ジクロロメタンで抽出し、得られた有機層を飽和食塩水で洗浄した後、無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、(S)-3-(((S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-メトキシ-3-オキソプロポキシ)メチル)モルホリン-4-カルボン酸アリル(4.5 g、37%)を得た。
窒素雰囲気下、塩化カルシウム(15.87 g、142.974 mmol)の水溶液(50 mL)に室温にて水酸化リチウム一水和物(0.91 g、37.999 mmol)を加え、室温で10分間撹拌した。反応液に上述の様にして得た(S)-3-(((S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-3-メトキシ-3-オキソプロポキシ)メチル)モルホリン-4-カルボン酸アリル(5.0 g、9.532 mmol)のテトラヒドロフラン溶液(50 mL)、イソプロパノール(200 mL)を0℃にて加え、室温にて9時間撹拌した後、1M塩酸水溶液を加え、減圧下溶媒を留去した。得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)にて精製することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(((S)-4-((アリルオキシ)カルボニル)モルホリン-3-イル)メチル)-L-セリン(化合物aa141、Fmoc-Ser(S-(Alloc)Mor-3-Me)-OH)(2.4 g、49%)を得た。
LCMS(ESI)m/z = 511 (M+H)+
保持時間:0.77分(分析条件SQDFA05)
ペプチド合成機によるペプチド合成に用いるレジンは、以下のとおり合成した。2-クロロトリチルクロライドレジン(100-200mesh、1%DVB)は渡辺化学工業及びChem-Impex社から購入した。

なお、本明細書では、ポリマーやレジンと化合物が結合した場合、ポリマーやレジン部位を○にて表記する場合がある。また、レジン部位の反応点を明確にさせる目的で、○に接続させて反応部位の化学構造を表記させる場合がある。上の構造では、Fmoc-Asp(O-Trt(2-Cl)-resin)-pip(化合物pd01)において、レジン上の2-クロロトリチル基がAspの側鎖カルボン酸とエステル結合を介して結合している様子を示している。


窒素雰囲気下、前工程にて得られた(S)-4-(2-(アリルオキシ)-2-オキソエチル)-5-オキソオキサゾリジン-3-カルボン酸(9H-フルオレン-9-イル)メチル(化合物pd03)(5.0 g、12.3 mmol)とトリエチルシラン(Et3SiH)(4.3 g、37.0 mmol)のジクロロメタン/トリフルオロ酢酸(TFA)=1/1(80 mL/80 mL)の溶液を室温にて2日間撹拌した後、反応液を減圧濃縮した。得られた残渣に炭酸カリウム(K2CO3)水溶液を加え、石油エーテルで3回洗浄した後、塩酸を用いてpH3まで調整し、酢酸エチルを用いて3回抽出した。有機層をまとめて、無水硫酸ナトリウムにて乾燥し、ろ過、減圧濃縮し、得られた残渣を逆相シリカゲルカラムクロマトグラフィー(0.1%ギ酸水溶液/0.1%ギ酸アセトニトリル)にて精製し、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-4-(アリルオキシ)-4-オキソブタン酸(化合物pd04、Fmoc-MeAsp(OAl)-OH)(3.0 g、46%、2工程)を得た。
LCMS(ESI)m/z=410(M+H)+
保持時間:0.84分(分析条件SQDFA05)

LCMS(ESI)m/z=477(M+H)+
保持時間:1.32分(分析条件SMDmethod6)

LCMS(ESI)m/z=437(M+H)+
保持時間:1.10分(分析条件SMDmethod6)

得られた(S)-3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-4-オキソ-4-(ピペリジン-1-イル)ブタン酸-2-クロロトリチルレジン(化合物pd07、Fmoc-MeAsp(O-Trt(2-Cl)-resin)-pip)(10.7 mg)を反応容器に入れ、DMF(200 μL)、ピペリジン(200 μL)を加えて室温にて1時間振とうした。その後、反応混合液にDMF(1.6 mL)を加え、400 μLを取りだし、その吸光度(301.2 nm)を測定し(Shimadzu、UV-1600PC(セル長1.0 cm)を用いて測定)、(S)-3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-4-オキソ-4-(ピペリジン-1-イル)ブタン酸-2-クロロトリチルレジン(化合物pd07、Fmoc-MeAsp(O-Trt(2-Cl)-resin)-pip)のローディング量を0.416 mmol/gと算出した。
なお、同様に合成したローディング量が異なる別ロットについてもペプチド合成に使用した。


特別な記載がない限り、A01-A10、B01-B10、C01-C17、D01-D17、E01-E08、F01-F08、G01-G11、H01-H11、I01-I06、J01-J06、AP01-AP143、pd30-pd36、pd50-pd70、pd100-pd129、pd162、pd163、pd167、pd177-pd247、pd386-pd395、pd452-pd454、pd482-pd487のペプチド化合物の合成は実施例1の冒頭に記載した基本ルートで、以下の方法にて行った。
WO2013/100132に記載の方法にてペプチド合成機(Multipep RS;Intavis社製)を用いて、Fmoc法によりペプチド合成を行った。なお、操作の詳細な手順については合成機に付属のマニュアルに従った。
合成機にN末端がFmocで保護されたアスパラギン酸の側鎖カルボン酸部位が結合した2-クロロトリチルレジン(例えば(S)-3-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-4-オキソ-4-(ピペリジン-1-イル)ブタン酸-2-クロロトリチルレジン(化合物pd01、Fmoc-Asp(O-Trt(2-Cl)-resin)-pip))(1カラムあたり100 mg)と、各種Fmoc-アミノ酸(0.6 mol/L)と1-ヒドロキシ-7-アザベンゾトリアゾール(HOAt)もしくはoxyma(0.375 mol/L)のN-メチル-2-ピロリドン(NMP)溶液と、ジイソプロピルカルボジイミド(DIC)のN,N-ジメチルホルムアミド(DMF)溶液(10% v/v)をセットした。Fmoc-Thr(THP)-OH(化合物aa01)はNMP溶液においてoxymaと共存させ、さらにモレキュラシーブス4A1/8(和光純薬工業)もしくはモレキュラシーブス4A1/16(和光純薬工業)を添加してセットした。Fmoc-Thr(THP)-OH(化合物aa01)と同様にアミノ酸側鎖の保護基にTHP基を有するアミノ酸(例えば、Fmoc-Ser(EtOTHP)―OH(化合物aa22)など)もまたNMP溶液においてoxymaと共存させ、さらにモレキュラシーブス4A1/8(和光純薬工業)もしくはモレキュラシーブス4A1/16(和光純薬工業)を添加してセットした。使用するアミノ酸が塩酸塩などの塩を用いる場合、DIPEA/DMF=3/1の溶液をセットし、ペプチド伸長の際に使用するアミノ酸に対して、0.96等量分のDIPEAを上記のDMF溶液として別途添加した。また、側鎖に酸性官能基を有するアミノ酸を含むペプチド化合物においては、脱保護剤の残留に起因するペプチドの過剰伸長を抑えるため、ペプチド伸長反応時のレジン洗浄に0.08 Mトリエチルアミン塩酸塩のジクロロメタン溶液を用いた。
WO2013/100132に記載の方法にて、上記の方法によって得られた固相上に担持された鎖状ペプチドに対し、DCMを加えレジンを再膨潤させた後、レジンに2,2,2-トリフルオロエタノール(TFE)/DCM(1/1,v/v,2 mL)を加えて室温にて2時間振とうした。続いてチューブ内の溶液を合成用カラムでろ過することによりレジンを除き、残ったレジンをさらに2,2,2-トリフルオロエタノール(TFE)/DCM(1/1,v/v,1 mL)にて2回洗浄した。得られた全ての切り出し溶液を混合した溶液、あるいはその溶液をさらにジクロロエタン(DCE)(4 mL)で希釈したものを減圧下濃縮した。なお2,2,2-トリフルオロエタノール(TFE)/DCM(1/1,v/v,2 mL)による切り出し時には用いたレジン上のモル数(ローディング量(mmol/g)に使用したレジン量(通常は0.10g)をかけたもの)に対して1.8等量となる容量のDIPEAを添加することも出来る。
切り出し後に減圧下濃縮した残渣をDMF/DCM(1/1,v/v,8mL)に溶解した。0.5MのO-(7-アザ-1H-ベンゾトリアゾール-1-イル)-N,N,N,N-テトラメチルウロニウムヘキサフルオロホスフェート(HATU)/DMF溶液(用いたレジン上のモル数(ローディング量(mmol/g)に使用したレジン量(通常は0.10 g)をかけたもの)に対して1.5等量となる容量)と、DIPEA(用いたレジン上のモル数に対して1.8等量)を加え、室温にて2時間以上撹拌した。その後、減圧下溶媒を留去した。目的の環状ペプチドの生成はLCMS測定によって確認した。
調製した0.05Mの硫酸水素テトラメチルアンモニウム/1,1,1,3,3,3-ヘキサフルオロイソプロピルアルコール(HFIP)溶液(2%トリイソプロピルシラン(TIPS))を4 mL加え、残渣を溶解させた後、室温にて4時間以上撹拌した。一定時間静置した後、ジイソプロピルエチルアミン(DIPEA)(70μL)を加え、減圧下溶媒を留去した。
なお、側鎖にAlloc基を有するアミノ酸を含むペプチド(例えばNva(2-R-4-F2-Pyrro)、Ser(S-4-F2-Pyrro-Me)などを配列中に含むペプチド)は、塩化亜鉛(用いたレジン上のモル数に対して1.0等量)、テトラキス(トリフェニルフォスフィン)パラジウム(0)(Pd(PPh3)4)(用いたレジン上のモル数に対して1.0等量)、ポリ(メチルヒドロシロキサン)(PMHS、CAS#63148-57-2)(用いたレジン上のモル数に対して0.9 M)、テトラヒドロフラン(用いたレジン上のモル数に対して0.1 M)を用いて室温で3時間30分以上撹拌することでAlloc基の脱保護を行った後、反応液に調製した0.05Mの硫酸水素テトラメチルアンモニウム/1,1,1,3,3,3-ヘキサフルオロイソプロピルアルコール(HFIP)溶液(2%トリイソプロピルシラン(TIPS))を4 mL加え、室温にて4時間以上撹拌した。一定時間静置した後、ジイソプロピルエチルアミン(DIPEA)(70μL)を加え、減圧下溶媒を留去した。Alloc基の脱保護においては、上記のPMHSを用いる代わりにテトラメチルジシロキサン(TMDS、CAS#3277-26-7)(用いたレジン上のモル数に対して4.0等量)を用いることができる。
いずれの場合においても、減圧下、溶媒を留去した後、DMF、DMSO、もしくは20%メタノール水溶液を加え、不溶物をフィルターろ過にて取り除いた後、preparative-HPLCで精製した。
0.05Mの硫酸水素テトラメチルアンモニウム/HFIP溶液(2%TIPS)は、HFIP(11.66mL)、TIPS(0.24mL)、DCE(0.10mL)を混合した溶液から4mLを抜き取り、これに34.3mgの硫酸水素テトラブチルアンモニウムを溶解させて調製した。この溶液(0.05Mの硫酸水素テトラメチルアンモニウム/HFIP溶液(2%TIPS))はHFIPの代わりに別のフルオロアルコール、例えば2,2,2-トリフルオロエタノール(TFE)なども用いることができる。
2-1 側鎖にドナーを有するアミノ酸でドナー部分に分子内水素結合を有するものと有しないものの膜透過性の比較
側鎖にドナーを有するアミノ酸でドナー部分に分子内水素結合を有するものが有しないものよりも良好な膜透過性をもつことを確認するため、それぞれのアミノ酸を含有する2つの環状ペプチドを合成し、国際公開第2018/124162号に記載の改良法により膜透過性を比較した。上記のような2つの環状ペプチドの組み合わせを複数個合成、評価することで一般性を確認した。その結果、(1)Ser(EtOH)とNle(6-OH)、(2)Ser(1-CF3-EtOH)とHnl(7-F3-6-OH)、(3)Tyr(3-OMe)とTyr、(4)Ser(S-4-F2-Pyrro-Me)とNva(2-R-4-F2-Pyrro)、(5)Ser(NMe-Aca)とGln(Me)の組み合わせにおいて側鎖にドナーを有するアミノ酸でドナー部分に分子内水素結合を有するものを含有するペプチドの方が有しないものを含むペプチドよりも良好な膜透過性をもつことが確認できた。
上記の概念をもとに設計したSer(EtOH)もしくはNle(6-OH)を含む環状ペプチドを合成し、国際公開第2018/124162号に記載の改良法により膜透過性を比較した。Ser(EtOH)を含む配列の化合物番号がAXX(XXは数字を意味する)、Nle(6-OH)を含む配列の化合物番号がBXX(XXは数字を意味する)である。化合物番号A01とB01がそれぞれ対応する化合物であり、以下同様にAXX、BXXとXXの部分が同じ配列同士がそれぞれ対応している組み合わせである。10個の組み合わせを比較したところ、全ての組み合わせでSer(EtOH)を含有する環状ペプチドの方がNle(6-OH)を含有する環状ペプチドよりも膜透過性に優れていた。
以下、特に記載のない限り同様の方法で解析した。
なお、A01を例にとり、表4、表6、表8、表10、表12および表14のカラムについて説明する。A01は11残基からなる環状ペプチドであり、11残基のアミノ酸で環を形成している。環化部分を1番目のアミノ酸と定義し、A01の場合ではAspが該当する。Aspの隣で環状部を構成するアミノ酸を2番目のアミノ酸と定義し(A01の場合MeLeuが該当)、以下3番目、4番目とN末端まで繰り返す(A01の場合3番目MePhe、4番目Ser(iPen)となり、環状部が11残基からなるためN末端は11番目のD-Val)。またC-termのカラムはC末端のアミノ酸のカルボン酸部位と縮合する官能基を意味しており、pipはピペリジンを意味している。そして、N末端のアミノ基と環化部のアミノ酸の側鎖カルボン酸をアミド結合により環化させたペプチドであるということを表4、表6、表8、表10、表12および表14で表している。例えば、A01はN末端であるD-Valのアミノ基と環化部のAspの側鎖カルボン酸がアミド結合により環化している。本文中に記載の環状ペプチドに関しては特に記述が無い限り、全て同様の手法で表現している。
Ser(1-CF3-EtOH)を含む配列の化合物番号がCXX、Hnl(7-F3-6-OH)を含む配列の化合物番号がDXXである。17個の組み合わせを比較したところ、15個の組み合わせでSer(1-CF3-EtOH)を含有する環状ペプチドの方がHnl(7-F3-6-OH)を含有する環状ペプチドよりも膜透過性に優れていた。
Tyr(3-OMe)を含む配列の化合物番号がEXX、Tyrを含む配列の化合物番号がFXXである。8個の組み合わせを比較したところ、全ての組み合わせでTyr(3-OMe)を含有する環状ペプチドの方がTyrを含有する環状ペプチドよりも膜透過性に優れていた。
Ser(S-4-F2-Pyrro-Me)を含む配列の化合物番号がGXX、Nva(2-R-4-F2-Pyrro)を含む配列の化合物番号がHXXである。11個の組み合わせを比較したところ、10個の組み合わせでSer(S-4-F2-Pyrro-Me)を含有する環状ペプチドの方がNva(2-R-4-F2-Pyrro)を含有する環状ペプチドよりも膜透過性に優れていた。
Ser(NMe-Aca)を含む配列の化合物番号がIXX、Gln(Me)を含む配列の化合物番号がJXXである。6個の組み合わせを比較したところ、全ての組み合わせでSer(NtBu-Aca)を含有する環状ペプチドの方がGln(Me)を含有する環状ペプチドよりも膜透過性に優れていた。


Ser(tBuOH)、Ser(NtBu-Aca)、bAla(3R-MeOEtOH)、Phe(3-OMe-4-CONMe)、Phe(4-OMe-3-CONMe)、Ser(3-Me-5-Oxo-Odz)、Ser(2-Me-2-BuOH)、Ser(S-2-PrOH)、Ser(R-2-PrOH)、Ser(nPrOH)、Ser(S-2-BuOH)、Ser(R-2-BuOH)、Ser(2-Me2-PrOH)、bAla(2S-MeOEtOH)、Phe(3-OMe-4-CONHMs)、Hph(3-OMe-4-CONMe)、D-Hph(3-OMe-4-CONMe)、(Ph(3-OMe-4-CONMe)Et)Gly、MeHph(3-OMe-4-CONMe)、Hyp(2-EtOH)、Ser(S-Mor-3-Me)、Phe(4-CONMeOH)、Phe(3-OMe-4-CONHOH)の膜透過性を評価した。結果は下記の通りであった。共に擬似6員環を形成するSer(nPrOH)とSer(2-Me2-PrOH)の膜透過性を比較するとジェミナルにメチル基が存在するSer(2-Me2-PrOH)の膜透過性の方がジェミナルにメチル基を有さないSer(nPrOH)よりも良好な傾向がみられた。この理由としてソープ・インゴールド効果による配座自由度の減少やドナー部分付近にジメチル基のような立体的に大きな脂溶性の置換基が存在することで極性表面積(Polar Surface Area:PSA)が減少しているなどといったことが考えられる。


実施例3-1 側鎖部分構造のLow energy conformationの計算方法
側鎖部分構造のLow energy conformationは側鎖β位(主鎖に直結する炭素)からアミノ酸の側鎖部分を抜き出した部分構造を用いて計算した。下スキームにSer(EtOH)、bAla(3R-MeOEtOH)、bAla(2S-MeOEtOH)、Phe(3-OMe-4-CONMe)の例を示す。


(1)初期構造発生
回転可能な単結合に関して初期二面角を0、90、180、270度と与えた。アミド結合ならびにCとOHの間の結合に関して初期二面角を0、180度と与えた。Ser(EtOH)の側鎖の部分構造を計算する場合、2個の単結合に4個づつ初期二面角、CとOHの間の結合に2個の初期二面角を考慮するので、合計32個の初期コンフォメーションを発生した。
(2)構造最適化
Gaussian09を用いて(1)で発生させたすべてのコンフォメーションの構造最適化を行った。B3LYP/6-31G(d) レベルにPolarizable Continuum Model (PCM)を加えて疑似水中での構造最適化を行った。
(3)Low energy conformationの選択

1)下記部分構造(例えばSer(NMe-Aca)の側鎖部分構造)でのポテンシャルエネルギー曲面(図3-1)の場合

(2)各コンフォメーション(grid point)のエネルギー計算
ΔE(φ2、φ3)=E(φ2、φ3)―Elowest
Elowest : lowest energy conformationのエネルギー

(2)各コンフォメーション(grid point)のエネルギー計算
φ2を初期値に固定して、Gaussian09を用いて発生させたすべてのコンフォメーションの構造最適化を行った。B3LYP/6-31G(d) レベルにPolarizable Continuum Model (PCM)を加えて疑似水中での構造最適化を行った。その結果、各φ2に対して4x2=8個のE(φ2)が得られる。このE(φ2)の中で最もエネルギーの低いものをElowest(φ2)とした。φ2を15°間隔で計算した24個のコンフォメーションのエネルギー(ΔE(φ2))は下記の式で求めた。ΔE(φ2)をプロットしたものが図4-1である。
ΔE(φ2)=Elowest(φ2)―Elowest
Elowest : lowest energy conformationのエネルギー
CSD(The Cambridge Structural Database)5.39+2 updatesを用いてqueryを以下のものとして検索した。

下記に記載のいずれアミノ酸においても、側鎖部分構造は分子内水素結合を有するコンフォメーションを取りやすい構造であることが計算的に示された。
Ser(EtOH)、bAla(3R-MeOEtOH)、bAla(2S-MeOEtOH)、Hyp(2-EtOH):これら3つのアミノ酸は側鎖部分構造は同一になる。計算結果は分子内水素結合を有するコンフォメーションが最安定コンフォメーションであった。
Hph(3-OMe-4-CONMe)-OH、D-Hph(3-OMe-4-CONMe)-OH、MeHph(3-OMe-4-CONMe)-OH、(Ph(3-OMe-4-CONMe)Et)Gly:これら4つのアミノ酸は側鎖部分構造は同一になる。計算結果は分子内水素結合を有するコンフォメーションが最安定コンフォメーションであった。
表17の計算結果のカラムは最安定コンフォメーションのみを記載した。


本明細書におけるアミノ酸の側鎖には、アミノ酸に含まれる炭素原子(例えば、α、β、又はγ-炭素原子)に結合している鎖、及び窒素原子に結合している鎖が含まれる。本明細書において、アミノ酸の側鎖の長さは、以下の方法により求めることができる。すなわち、アミノ酸単位のN末をアセチル基、C末をメチルアミノ基でキャッピングし、分子モデリングソフトMOE(Chemical Computing Group社)のLowModeMDにより配座を発生させ、側鎖部位が結合している原子(天然アミノ酸の場合はα-炭素原子(Cα炭素))から最も遠い同側鎖の原子(水素原子は除く)までの距離を測定することで求めることができる。例としてPheまたはnBuGlyの側鎖の長さの求め方を以下に示す。


本明細書においてアミノ酸の側鎖、又は環状ペプチド化合物の環状部の側鎖の計算pKa及びBasic 計算pKaは、ADMETPredictor(Simulations Plus Inc.社、ver.8.0)を用いて求めることができる。計算pKa及びBasic 計算pKaは側鎖β位(主鎖に直結する炭素)から側鎖部分を抜き出した部分構造を用いて計算される。例としてLysの場合を以下に示す。側鎖β位(主鎖に直結する炭素)を含む部分構造を用いてBasic 計算pKaを計算すると10.5であった。同様に計算すると、酸としては、Aspの側鎖力ルボキシ基の計算pKaが4.3であり、Tyrの側差フェノール性水酸基の計算pKaが9.9であり、3-フルオロチロシン(Tyr(3-F))の側差フェノール性水酸基の計算pKaが8.7であり、テトラゾールの計算pKaが3.7であった。一方、塩基としては、Argの側鎖グアニジノ基のBasic 計算pKaが12.7であり、Hisのイミダゾリル基のBasic 計算pKaが7.6であり、ピリジンのBasic 計算pKaが5.4であった。


本明細書においてアミノ酸側鎖のpKaの実測は以下の手順に従って行った。
(使用装置)
Sirius T3(Sirius Analytical Instruments Ltd., Forest Row, East Sussex, RH18 5DW, UK)

被験物質約1mgの粉体または10mMのDMSO溶液約100uLに0.15MのKClを添加し、塩基性化合物の場合はpH2となるまで0.5MのHClを添加後、0.5MのKOHにてpH12まで滴定を行った。
酸性化合物の場合は、pH12となるまで0.5MのKOH添加後、0.5MのHClにてpH2まで滴定を行った。
滴定操作はSirius社製T3にて自動で行い、得られた滴定曲線からsirius T3 Refinementソフトを用いてpKaを求めた。
また参照化合物としてイミプラミン・HCl、プロプラノロール・HCl、ワルファリンを用いて同様にpKaを求め、Sirius社で得られた試験結果範囲内であることを確認した。
アミノ酸側鎖のpKaは評価したいアミノ酸を下記に示す3残基からなるペプチドの2残基目(下記表のXの位置)に導入した配列を合成することで求めた。


実施例1に記載のペプチド化合物の化学合成と同様の手法を用いペプチドの伸長、レジン上からの切り出しを行った後、C末端のカルボン酸とピペリジンを縮合することでpd30-pd37を合成した。また、ペプチド合成に用いたFmoc-MeGly-Trt(2-Cl)-resin(化合物pd11)は下記の通り合成した。ZMeGly(Cbz-MeGly-OH、CAS#39608-31-6)は東京化成工業より購入した。

得られたレジンのローディング量は文献記載(Letters in Peptie Science, 2002, 9, 203)の方法を用い算出した。N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチルグリシン-2-クロロトリチルレジン(化合物pd11、Fmoc-MeGly-Trt(2-Cl)-resin)(14.4mg)を反応容器に入れ、DMF(2mL)を加え1時間振とうした。反応液にDBU(0.04mL)を加え、30分振とうした後、DMF(10mL)を加え、1mLを取りだし、さらに溶液量が12.5mLになるまでDMFで希釈した。得られた溶液の吸光度(294nm)を測定し(Shimadzu、UV-1600PC(セル長1.0cm)を用いて測定)、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N-メチルグリシン-2-クロロトリチルレジン(化合物pd11、Fmoc-MeGly-Trt(2-Cl)-resin)のローディング量を0.789mmol/gと算出した。


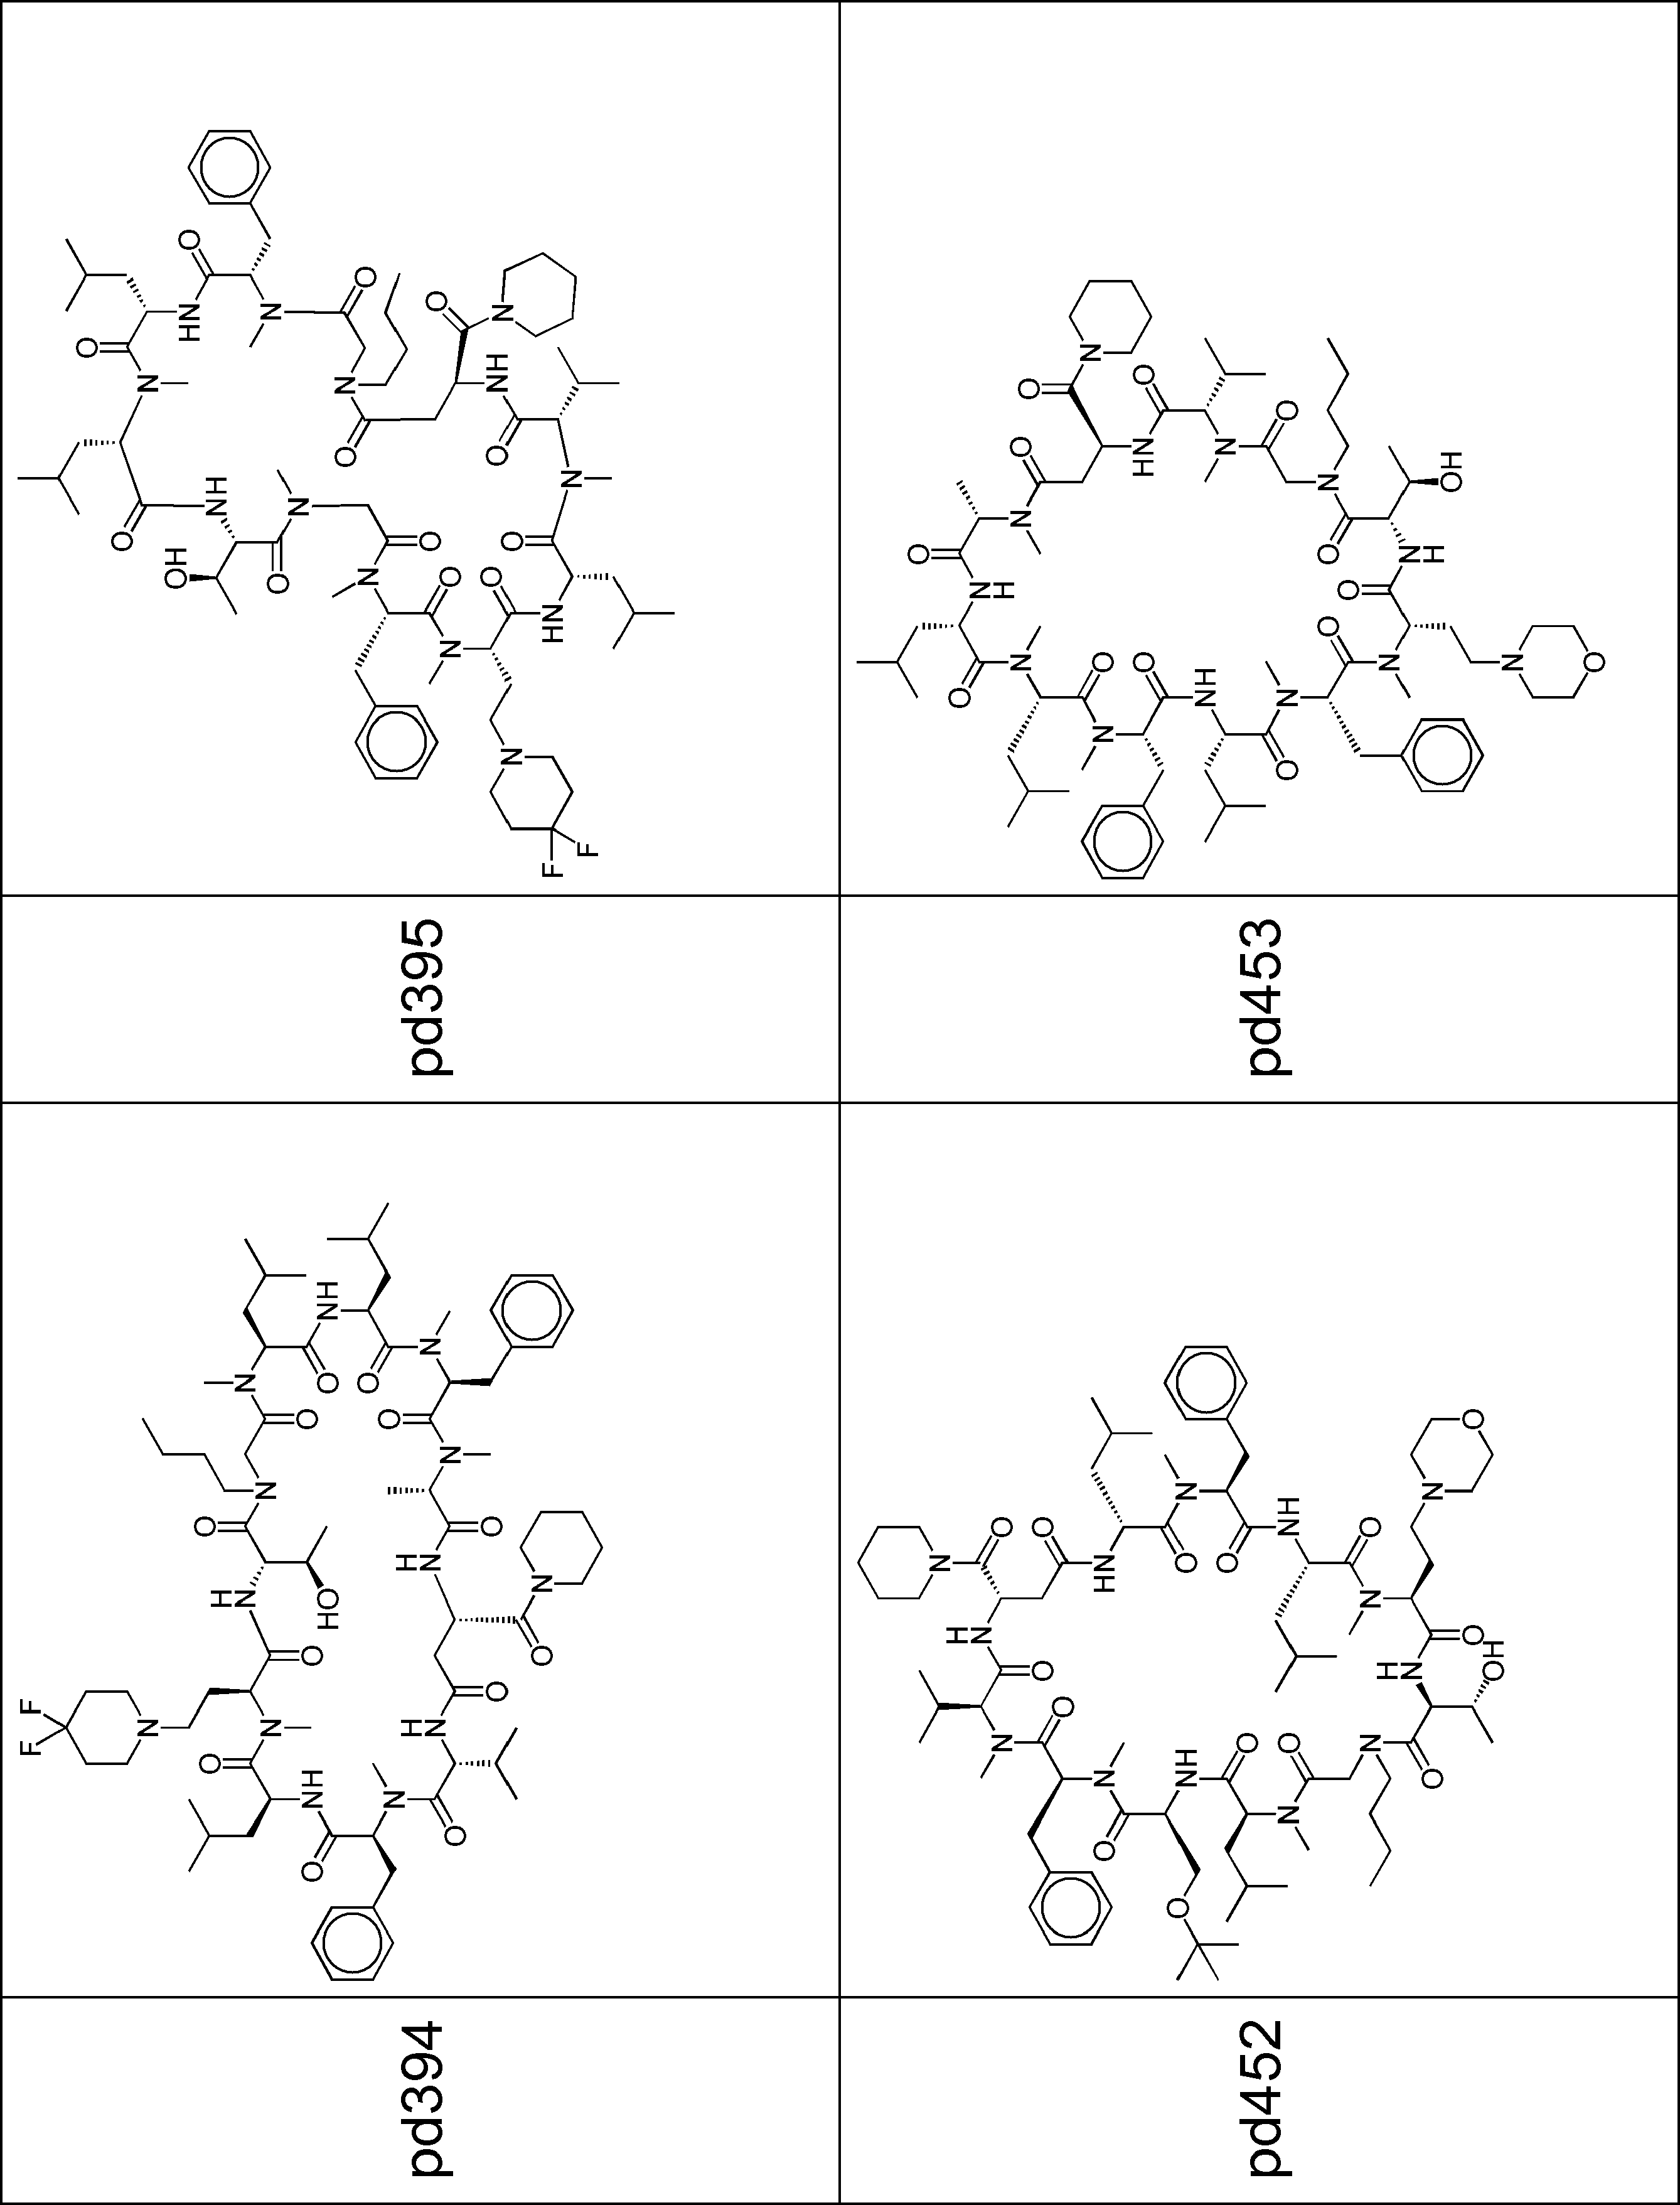


pKa実測に用いた3残基ペプチドおよび膜透過性を評価した環状ペプチドを合成するために使用したFmoc-Ser(Et-2-NMe2)-OH(化合物tm01)、Fmoc―MeAbu(pip-4-F2)-OH(化合物tm04)、Fmoc―MeAbu(pip-3-F2)-OH(化合物tm10)、Fmoc-MeAbu(Mor)-OH(化合物tm19)、Fmoc-Ser(Et-2-Mor)-OH(化合物tm20)、Fmoc-Abu(5-Oxo-Odz)-OH(化合物tm21)、Fmoc-Gln(Ms)-OH(化合物tm22)は以下のように合成した。

得られた、O-(2-(ジメチルアミノ)エチル)-N-トリチル-L-セリン(Trt-Ser(Et-2-NMe2)-OH)(4.1g、9.80mmol)にジクロロメタン(10mL)を加えた後、4N塩酸/1,4-ジオキサン溶液(40mL、160mmol)、水(4mL)を加え、室温で3時間撹拌した。反応液に水(80mL)を加え、ヘキサンで2回洗浄し、得られた水層に0度にて炭酸ナトリウム(26.0g、245mmol)、1、4-ジオキサン(120mL)、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(3.97g、11.76mmol)を加え、室温にて2時間撹拌した。反応液にギ酸(11.28mL、294mmol)を加えた後、減圧下1,4-ジオキサンを留去し、得られた水層を逆相カラムクロマトグラフィー(0.1%ギ酸水溶液/0.1%ギ酸アセトニトリル溶液)で精製することでN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-(ジメチルアミノ)エチル)-L-セリン(化合物tm01、Fmoc-Ser(Et-2-NMe2)-OH)のギ酸塩(3.07g、3工程69%)を得た。得られたN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-(ジメチルアミノ)エチル)-L-セリン(化合物tm01、Fmoc-Ser(Et-2-NMe2)-OH)のギ酸塩(3.0g、7.53mmol)にジクロロメタン(4mL)、4N塩酸/1、4-ジオキサン溶液(9.411mL、37.6mmol)を加え、室温にて20分撹拌した。反応液を減圧濃縮することで、N-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-(ジメチルアミノ)エチル)-L-セリン(化合物tm01、Fmoc-Ser(Et-2-NMe2)-OH)の塩酸塩(2.7g、82%)を得た。
LCMS(ESI) m/z = 399 (M+H)+
保持時間:0.50分(分析条件SQDFA05)

LCMS(ESI)m/z = 515 (M+H)+
保持時間:0.97分(分析条件SQDFA05)

LCMS(ESI)m/z = 445 (M+H)+
保持時間:0.56分(分析条件SQDFA05)

LCMS(ESI)m/z = 459 (M+H)+
保持時間:2.71分(分析条件SMD method28)

LCMS(ESI)m/z = 212 (M+H)+
保持時間:0.94分(分析条件SMD method31)

別途合成した、(S)-2-(((ベンジルオキシ)カルボニル)(メチル)アミノ)-4-(エチルチオ)-4-オキソブタン酸ベンジル(化合物tm08、Cbz-MeAsp(SEt)-OBn)(20g、48.13mmol)、3,3-ジフルオロピペリジン(化合物tm06)(8.26g、68.19mmol)、トリエチルシラン(40mL)、10%パラジウム/炭素(2.6g)のN,N-ジメチルホルムアミド溶液(20mL)に0度にてナトリウムトリアセトキシボロヒドリド(NaBH(OAc)3)(20.4g、204mmol)を加え、室温にて30分間撹拌した。反応液をろ過、減圧濃縮することで得た粗生成物を順相シリカゲルクロマトグラフィー(ヘキサン/酢酸エチル)にて精製し、(S)-2-(((ベンジルオキシ)カルボニル)(メチル)アミノ)-4-(3,3-ジフルオロピペリジン-1-イル)ブタン酸ベンジル(化合物tm07、Cbz-MeAbu(pip-3-F2)-OBn)(11g、50%)を得た。
LCMS(ESI)m/z = 461 (M+H)+
保持時間:1.42分(分析条件SMD method32)

上述の粗生成物である(2S)-4-(3,3-ジフルオロピペリジン-1-イル)-2-(メチルアミノ)ブタン酸の臭化物(化合物tm09)(7.08g、29.97mmol)とN-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(16g、47.48mmol)を炭酸カリウムの水/1,4-ジオキサン溶液=1/1(300mL)に溶解し室温にて4時間撹拌した。その後反応液をジエチルエーテルにて3回洗浄し、塩酸で水層のpHを3に調節し、酢酸エチルにて2度抽出した。得られた有機層を合わせ、無水硫酸ナトリウムで乾燥、ろ過、減圧濃縮し、得られた残渣を逆相カラムクロマトグラフィー(10mmol炭酸水素アンモニウム水溶液/アセトニトリル)にて精製し、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)(メチル)アミノ)-4-(3,3-ジフルオロピペリジン-1-イル)ブタン酸(化合物tm10、Fmoc-MeAbu(pip-3-F2)-OH))(7.2g、71%)を得た。
LCMS(ESI)m/z = 459 (M+H)+
保持時間:1.61分(分析条件SMD method5)

上述と同様の方法で合成した(S)-2-(3-((ベンジルオキシ)カルボニル)-5-オキソオキサゾリジン-4-イル)酢酸(化合物tm11)(265g、948.99mmol)の粗生成物とエタンチオール(88.3g、1.42mol)と4-ジメチルアミノピリジン(DMAP)(11.59g、95mmol)のジクロロメタン溶液に0度にてN,N’-ジシクロヘキシルカルボジイミド(DCC)(214g、1.04mol)を加え、室温にて5時間撹拌した。反応溶液をろ過し、得られたろ液を飽和食塩水で洗浄し、得られた有機層を無水硫酸ナトリウムで乾燥させた後、ろ過、減圧濃縮した。得られた粗生成物を順相シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル)にて精製し、(S)-4-(2-(エチルチオ)-2-オキソエチル)-5-オキソオキサゾリジン-3-カルボン酸ベンジル(化合物tm12)(167g)の混合物を得た。
得られた(S)-4-(2-(エチルチオ)-2-オキソエチル)-5-オキソオキサゾリジン-3-カルボン酸ベンジル(化合物tm12)(139g、1.20mol)のジクロロメタン(DCM)とトリフルオロ酢酸溶液(1500mL/1500mL)にトリエチルシラン(139g、1.2mol)を加え、室温にて3日間撹拌した。反応液を減圧濃縮し得られた残渣に炭酸カリウム水溶液を加え、ジエチルエーテルで3度洗浄した。得られた水層を2N塩酸水溶液でpHを3に調節したものに対して酢酸エチルで2度抽出した。合わせた有機層を飽和食塩水で洗浄し、有機層を無水硫酸ナトリウムで乾燥させた後、ろ過、減圧濃縮し、(S)-2-(((ベンジルオキシ)カルボニル)(メチル)アミノ)-4-(エチルチオ)-4-オキソブタン酸(化合物tm13、Cbz-MeAsp(SEt)-OH)(75g)の粗生成物を得た。
LCMS(ESI)m/z = 326 (M+H)+
保持時間:1.00分(分析条件SMD method16)

1HNMR(300MHz,CDCl3):δ7.36-7.31(m,10H),5.19-4.89(m,5H),3.34-2.85(m,7H),1.30-1.21(m,3H)


LCMS(ESI) m/z = 478 (M+Na)+
保持時間:3.13分(分析条件 SMD method14)

LCMS(ESI) m/z = 418 (M+Na)+
保持時間:2.07分(分析条件 SMD method40)

LCMS(ESI) m/z = 467 (M+H)+
保持時間:2.02分(分析条件 SMD method41)

LCMS(ESI) m/z = 411 (M+H)+
保持時間:1.20分(分析条件 SMD method42)


LCMS(ESI) m/z = 423 (M+H)+
保持時間:1.14分(分析条件 SMD method40)

LCMS(ESI) m/z = 425 (M+H)+
保持時間:1.40分(分析条件 SMD method34)

N-(tert-ブトキシカルボニル)-O-(2-モルホリノエチル)-L-セリン(Boc-Ser(Et-2-Mor)-OH)(5.6g、17.59mmol)に4N塩酸/1,4-ジオキサン溶液(28.0mL、921.53mmol)を室温にて加え、1時間撹拌した。減圧下溶媒を留去し、得られた残渣をジクロロメタンで洗浄した後、水(10mL)を加え溶液とした。得られた溶液に炭酸カリウムを加えpH7に調節し、水(100mL)と1.4-ジオキサン(150mL)を加えた後、室温にて炭酸カリウム(4.465g、32.07mmol)、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(5.68g、16.84mmol)を加え、16時間撹拌した。反応液をジエチルエーテルで洗浄し、濃塩酸を用いて水層のpHを1に調節し、ジクロロメタンで3回抽出した。得られた有機層を水で洗浄し、無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(0.1%塩酸水溶液/0.1%塩酸アセトニトリル溶液)で精製することでN-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-O-(2-モルホリノエチル)-L-セリン(化合物tm20、Fmoc-Ser(Et-2-Mor)-OH)の塩酸塩(3.2g、2工程48%)を得た。
LCMS(ESI) m/z = 441 (M+H)+
保持時間:0.50分(分析条件SQDFA05)

上述のように得られた(S)-2-((tert-ブトキシカルボニル)アミノ)-4-シアノブタン酸(50g、219.06mmol)のエタノール溶液(500mL)に室温にてヒドロキシルアミンの塩酸塩(32g、460.50mmol)とトリエチルアミン(83mL)を加え、80度で2時間撹拌した。反応液を減圧下濃縮することで、(S)-2-((tert-ブトキシカルボニル)アミノ)-5-(ヒドロキシアミノ)-5-イミノペンタン酸(107g)を粗生成物として得た。
窒素雰囲気下、上述の粗生成物である(S)-2-((tert-ブトキシカルボニル)アミノ)-5-(ヒドロキシアミノ)-5-イミノペンタン酸(44g、168.40mmol)の1,4-ジオキサン溶液(500mL)に室温にて1,1’-カルボニルジイミダゾール(CDI)(39g、240.52mmol)と1,8-ジアザビシクロ[5.4.0]-7-ウンデセン(DBU)(55g、361.27mmol)を加え、110度で4時間撹拌した。反応液を室温にて冷却した後、濃塩酸を加えpH=2になるように調整し、ジクロロメタンで2回抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過を行った後、減圧下溶媒留去することで、(S)-2-((tert-ブトキシカルボニル)アミノ)-4-(5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)ブタン酸(Boc-Abu(5-Oxo-Odz)-OH)(11g)を粗生成物として得た。
窒素雰囲気下、上述のように得られた(S)-2-((tert-ブトキシカルボニル)アミノ)-4-(5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)ブタン酸(Boc-Abu(5-Oxo-Odz)-OH)(13.5g、46.99mmol)の1,4-ジオキサン溶液(10mL)に室温にて4N塩酸/1,4-ジオキサン溶液(140mL)を加え、16時間撹拌した。反応液を減圧下濃縮することで、(S)-2-アミノ-4-(5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)ブタン酸(H-Abu(5-Oxo-Odz)-OH)(8.8g)を粗生成物として得た。
窒素雰囲気下、得られた粗生成物である(S)-2-アミノ-4-(5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)ブタン酸(H-Abu(5-Oxo-Odz)-OH)(8.8g、47.02mmol)と炭酸カリウム(13g、94.06mmol)の水/1,4-ジオキサン溶液(100mL/100mL)に室温にて、N-(9-フルオレニルメトキシカルボニルオキシ)スクシンイミド(Fmoc-OSu)(14.3g、42.4mmol)を加え、3時間撹拌した。反応液をt-ブチルメチルエーテル/ヘキサン(1/3)で洗浄し、濃塩酸を用いて水層のpHを2に調節し、ジクロロメタンで2回抽出した。得られた有機層を無水硫酸ナトリウムで乾燥、ろ過を行った。ろ液を減圧下濃縮し、得られた残渣を逆相カラムクロマトグラフィー(水/アセトニトリル)で精製することで、(S)-2-((((9H-フルオレン-9-イル)メトキシ)カルボニル)アミノ)-4-(5-オキソ-4,5-ジヒドロ-1,2,4-オキサジアゾール-3-イル)ブタン酸(化合物tm21、Fmoc-Abu(5-Oxo-Odz)-OH)(2.67g)を得た。
LCMS(ESI) m/z = 410 (M+H)+
保持時間:0.66分(分析条件SQDFA05)

窒素雰囲気下、上述のようにして得られた、tert-ブチルN2-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N5-(メチルスルホニル)-L-グルタミナート(Fmoc-Gln(Ms)-OtBu)(5.5g、10.9mmol)の2,2,2-トリフルオロエタノール(TFE)溶液(100mL)に0度にてクロロトリメチルシラン(TMSCl)(3.57g、32.861mmol))を加え、1時間撹拌した。反応液を減圧下濃縮し、得られた残渣にt-ブチルメチルエーテルを加え再び濃縮した。この操作をさらに2回繰り返し、アセトニトリル/ジクロロメタンで再結晶を行うことで、N2-(((9H-フルオレン-9-イル)メトキシ)カルボニル)-N5-(メチルスルホニル)-L-グルタミン(化合物tm22、Fmoc-Gln(Ms)-OH)(3.8178g、77%)を白色固体として得た。
LCMS(ESI) m/z = 447 (M+H)+
保持時間:0.65分(分析条件SQDFA05)
Trpの側鎖を有する環状ペプチド化合物、及び同側鎖を有しない環状ペプチド化合物を合成し、改良法により膜透過性試験を行った。なお、MeTrpはTrpの側鎖を含むことから、構成アミノ酸にMeTrpを含む環状ペプチド化合物も、Trpの側鎖を有する環状ペプチド化合物に含まれる。その結果、Trpの側鎖を持たない環状ペプチド化合物はClogPを上昇させることによりPapp≧1.0×10⁻⁶cm/秒を達成することが可能であった(図1-1)。一方、Trpの側鎖を1つ以上有しているとClogPを上昇させてもPapp≧1.0×10-6cm/秒まで膜透過性を上げるのは難しかった(図1-2)。例えばTrp側鎖の位置、Nアルキルのパターンを変えたもので、1.06≦ClogP/total aa≦1.43の範囲にある環状ペプチド化合物の配列を表31に示すが、これらはPapp≧1.0×10⁻⁶cm/秒の膜透過性を達成することは難しかった。
以上のことから、Trpの側鎖を持たない環状ペプチド化合物の膜透過性を評価したところ、ClogPを上昇させることにより膜透過性が向上することが確認できたため(図1-1)、比較する2つのペプチドのClogP/total AAの値が±0.03の間となるように調節した。


Tyr(3-F)又はTyrの側鎖を有する環状ペプチド化合物を合成し、改良法により膜透過性試験を行った。その結果、Tyr(3-F)又はTyrの側鎖を有する環状ペプチド化合物もTrpの側鎖を有する環状ペプチド化合物と同様の傾向であった。1.06≦ClogP/total aa≦1.29の範囲にある環状ペプチド化合物の配列を表33に示すが、Trpの場合と同様、これらがPapp≧1.0×10⁻⁶cm/秒を達成することは困難であった。


ARCが増加するにともなって環状ペプチド化合物のPappが低下、すなわち膜透過性が低下する傾向が確認できた(図2)。そのため、比較する2つのペプチドの芳香環の数(Aromatic Ring Count: ARC)を揃えた。
なお、本解析では、環状部の側鎖に縮環構造を有するもの、並びに置換及び無置換のヒドロキシフェニル基を有する環状ペプチド化合物を解析対象から除外した。ARC=3の場合、ClogP/total AAが0.88以上の配列で、1.0×10⁻⁶以上のPappが認められた。一方、ARC=4の場合は前記ARC=3の化合物群と同程度のClogP/total AA(0.88以上)の化合物であっても、Pappが1.0×10⁻⁶以上となるものはほとんど存在しなかった。このことから、同程度の脂溶性の配列群であるにも関わらず、ARC=3とARC=4を比較すると、ARC=3の方が良好な膜透過性を示すことが示された(表35、表36、表37)。
また、pd198を例にとり、表35、表36及び表37のカラムについて説明する。pd198は12残基からなる環状ペプチドであり、10残基のアミノ酸で環を形成している。環化部分を1番目のアミノ酸と定義し、pd198の場合ではAspが該当する。Aspの隣で環状部を構成するアミノ酸を2番目のアミノ酸と定義し(pd198の場合Pheが該当)、以下3番目、4番目とN末端まで繰り返す(pd198の場合3番目MeVal、4番目MeLeuとなり、環状部が10残基からなるためN末端は10番目のg-MeAbu)。また環化部のアミノ酸のC末端から近い順に、H-1、H-2、H-3と示している。pd198の場合H-1はMePhe、H-2はAlaであるので、AspのC末端にMePheが結合しており、ついでAlaが結合していることを意味している。またC-termのカラムはC末端のアミノ酸のカルボン酸部位と縮合する官能基を意味しており、pipはピペリジンを表し、pyrroはピロリジンを意味している(pd198の場合、C末端のアミノ酸はH-2のAlaであり、そのカルボン酸部位がpip(ピペリジン)と縮合した構造になる)。そして、N末端のアミノ基と環化部のアミノ酸の側鎖カルボン酸をアミド結合により環化させたペプチドであるということを表35、表36及び表37で表している。例えば、pd198はN末端であるg-MeAbuのアミノ基と環化部のAspの側鎖カルボン酸がアミド結合により環化している。本文中に記載の環状ペプチドに関しては特に記述が無い限り、全て同様の手法で表現している。


Claims (16)
- 2以上のアミノ酸が連結されたペプチド化合物であって、アミノ酸の少なくとも1つがその側鎖内で水素結合を形成し得る、ペプチド化合物。
- 側鎖内で水素結合を形成し得るアミノ酸が、側鎖内で擬似4~7員環を形成し得る、請求項1に記載のペプチド化合物。
- 側鎖内で水素結合を形成し得るアミノ酸が、下記式Aで表される、請求項1または2に記載のペプチド化合物:
式中、
R1は、水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、この場合、R2AおよびR2Bは、R2Aが水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、かつR2Bは、水素またはC1-C6アルキルであるか、あるいは、R2AおよびR2Bは、それらが結合している炭素原子と一緒になって4~6員の環を形成し、あるいは
R1は、R1が結合する窒素原子、R2A、およびR2Aが結合する炭素原子と一緒になって4~6員の複素環を形成し、該複素環は、式1もしくは式2で表される基、-OH、およびアルコキシ基からなる群より選択される、1つまたは複数の置換基を有していてもよく、この場合、R2Bは、水素またはC1-C6アルキルであり、
R3は、単結合、または-CHR4-であり、
R4は、水素、もしくはC1-C4アルキルであるか、または式1もしくは式2で表される基であり、
式1および式2は、それぞれ下記式:
で表され、式中、
*は、結合点を示し、
Q1およびQ2は、独立して、単結合、C1-C4アルキレン、または酸素原子を1つ含むC2-C4ヘテロアルキレンであり、
A1は、-O-、または-S-であり、
L1は、フッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
A2は、単結合、-O-、または-S-であり、
L2は、単結合であるか、またはフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xは、-OH、-NRZ1RZ2、もしくは-CONRZ1RZ2であるか、または1~3個のヘテロ原子を含有し、オキソもしくは1つまたは複数のハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり;
RZ1およびRZ2は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Yは、-OH、C1-C4アルキルスルホニルアミノ、-NRZ3RZ4、もしくは-CONRZ3RZ4であるか、または1~3個のヘテロ原子を含有し、オキソもしくはハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり、
RZ3およびRZ4は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Zは、水素またはC1-C4アルキルであり、
但し、A2が-O-、または-S-である場合、Zは水素ではなく、
ここで、式Aで表されるアミノ酸は、式1または式2で表される基のいずれかを少なくとも1つ含む。 - 式Aで表されるアミノ酸が以下からなる群:

 より選択される、請求項3に記載のペプチド化合物。

- 5~30のアミノ酸から構成される、請求項1~4のいずれか1項に記載のペプチド化合物。
- 環状である、請求項1~5のいずれか1項に記載のペプチド化合物。
- 2~15のアミノ酸から構成される環状部を含む、請求項6に記載のペプチド化合物。
- 2~30のN置換アミノ酸を含む、請求項1~7のいずれか1項に記載のペプチド化合物。
- 全アミノ酸数に対するN置換アミノ酸数の割合が30%以上である、請求項1~8のいずれか1項に記載のペプチド化合物。
- 4.0~18のClogPを有する、請求項1~9のいずれか1項に記載のペプチド化合物。
- 1.0x10⁻⁷cm/秒以上のPappを有する、請求項1~10のいずれか1項に記載のペプチド化合物。
- 請求項1~11のいずれか1項に記載のペプチドおよび/または当該ペプチドをコードする核酸を含むライブラリ。
- 下記式Aで表される、アミノ酸:
式中、
R1は、水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、この場合、R2AおよびR2Bは、R2Aが水素、もしくはC1-C6アルキルであるか、または式1もしくは式2で表される基であり、かつR2Bは、水素またはC1-C6アルキルであるか、あるいは、R2AおよびR2Bは、それらが結合している炭素原子と一緒になって4~6員の環を形成し、あるいは
R1は、R1が結合する窒素原子、R2A、およびR2Aが結合する炭素原子と一緒になって4~6員の複素環を形成し、該複素環は、式1もしくは式2で表される基、-OH、およびアルコキシ基からなる群より選択される、1つまたは複数の置換基を有していてもよく、この場合、R2Bは、水素またはC1-C6アルキルであり、
R3は、単結合、または-CHR4-であり、
R4は、水素、もしくはC1-C4アルキルであるか、または式1もしくは式2で表される基であり、
式1および式2は、それぞれ下記式:
で表され、式中、
*は、結合点を示し、
Q1およびQ2は、独立して、単結合、C1-C4アルキレン、または酸素原子を1つ含むC2-C4ヘテロアルキレンであり、
A1は、-O-、または-S-であり、
L1は、フッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
A2は、単結合、-O-、または-S-であり、
L2は、単結合であるか、またはフッ素、C1-C2アルキル基、C1-C2フルオロアルキル、およびオキソ(=O)からなる群より選択される1つまたは複数の置換基によって置換されていてもよい直鎖のC1-C3アルキレンであり、
Xは、-OH、-NRZ1RZ2、もしくは-CONRZ1RZ2であるか、または1~3個のヘテロ原子を含有し、オキソもしくは1つまたは複数のハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり;
RZ1およびRZ2は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Yは、-OH、C1-C4アルキルスルホニルアミノ、-NRZ3RZ4、もしくは-CONRZ3RZ4であるか、または1~3個のヘテロ原子を含有し、オキソもしくはハロゲンによって置換されていてもよい5~6員の飽和もしくは不飽和ヘテロシクリルであり、
RZ3およびRZ4は、独立して、水素、-OH、C1-C4アルキル、およびC1-C4アルキルスルホニルからなる群より選択され、
Zは、水素またはC1-C4アルキルであり、
但し、A2が-O-、または-S-である場合、Zは水素ではなく、
ここで、式Aで表されるアミノ酸は、式1または式2で表される基のいずれかを少なくとも1つ含む。 - 以下からなる群:


 より選択される、請求項13に記載のアミノ酸。

- 請求項13または14に記載のアミノ酸に含まれるアミノ基、および/またはカルボキシル基が保護基で保護されている、保護アミノ酸。
- アミノ基の保護基が、Fmoc基、Boc基、Cbz基、Alloc基、ノシル基、ジニトロノシル基、t-Bu基、トリチル基、およびクミル基からなる群より選択される、および/または
カルボキシル基の保護基が、メチル基、アリル基、t-Bu基、トリチル基、クミル基、メトキシトリチル基、およびベンジル基からなる群より選択される、請求項15に記載の保護アミノ酸。
Priority Applications (8)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| CN201980091739.9A CN113454058B (zh) | 2018-12-12 | 2019-12-12 | 具有能形成分子内氢键的官能团的氨基酸、包含该氨基酸的肽化合物、及其制备方法 |
| JP2020559315A JP7531401B2 (ja) | 2018-12-12 | 2019-12-12 | 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 |
| EP19896830.7A EP3896056A4 (en) | 2018-12-12 | 2019-12-12 | AMINO ACID WITH FUNCTIONAL GROUP FOR INTERMOLECULAR HYDROGEN BONDING, PEPTIDE COMBINATION THEREOF AND PROCESS FOR THEIR PREPARATION |
| US17/312,296 US20230096766A1 (en) | 2018-12-12 | 2019-12-12 | Amino acid having functional group capable of intermolecular hydrogen bonding, peptide compound containing same and method for production thereof |
| CN202410282724.3A CN118184532A (zh) | 2018-12-12 | 2019-12-12 | 具有能形成分子内氢键的官能团的氨基酸、包含该氨基酸的肽化合物、及其制备方法 |
| KR1020217021220A KR20210102326A (ko) | 2018-12-12 | 2019-12-12 | 분자내 수소 결합 가능한 작용기를 갖는 아미노산과 그들 아미노산을 포함하는 펩타이드 화합물, 및 그들의 제조 방법 |
| JP2024103888A JP7757469B2 (ja) | 2018-12-12 | 2024-06-27 | 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 |
| JP2025170035A JP2025188158A (ja) | 2018-12-12 | 2025-10-08 | 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2018-232144 | 2018-12-12 | ||
| JP2018232144 | 2018-12-12 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2020122182A1 true WO2020122182A1 (ja) | 2020-06-18 |
Family
ID=71076440
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/JP2019/048720 Ceased WO2020122182A1 (ja) | 2018-12-12 | 2019-12-12 | 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 |
Country Status (6)
| Country | Link |
|---|---|
| US (1) | US20230096766A1 (ja) |
| EP (1) | EP3896056A4 (ja) |
| JP (3) | JP7531401B2 (ja) |
| KR (1) | KR20210102326A (ja) |
| CN (2) | CN113454058B (ja) |
| WO (1) | WO2020122182A1 (ja) |
Cited By (25)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2022138891A1 (ja) | 2020-12-25 | 2022-06-30 | 中外製薬株式会社 | N-置換-アミノ酸残基を含むペプチド化合物の製造方法 |
| WO2022145444A1 (ja) | 2020-12-28 | 2022-07-07 | 中外製薬株式会社 | アミノ酸の固相合成用樹脂への担持方法 |
| US11492369B2 (en) | 2017-12-15 | 2022-11-08 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide, and method for processing bases |
| JPWO2022234853A1 (ja) * | 2021-05-07 | 2022-11-10 | ||
| WO2022234850A1 (ja) | 2021-05-07 | 2022-11-10 | 中外製薬株式会社 | 環状ペプチド化合物を含む製剤及びその製造方法 |
| US11542299B2 (en) | 2017-06-09 | 2023-01-03 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptide containing N-substituted amino acid |
| WO2023127869A1 (ja) | 2021-12-28 | 2023-07-06 | 中外製薬株式会社 | N-アルキルアミノ酸、およびn-アルキルアミノ酸を含むペプチドの製造方法 |
| US11732002B2 (en) | 2018-11-30 | 2023-08-22 | Chugai Seiyaku Kabushiki Kaisha | Deprotection method and resin removal method in solid-phase reaction for peptide compound or amide compound, and method for producing peptide compound |
| US11891457B2 (en) | 2011-12-28 | 2024-02-06 | Chugai Seiyaku Kabushiki Kaisha | Peptide-compound cyclization method |
| WO2024043251A1 (ja) | 2022-08-23 | 2024-02-29 | 富士フイルム株式会社 | 環状ペプチドの細胞膜透過性の予測方法 |
| WO2024096023A1 (ja) | 2022-11-01 | 2024-05-10 | 中外製薬株式会社 | ジベンゾフルベンまたはジベンゾフルベン誘導体の除去方法 |
| WO2024143514A1 (ja) | 2022-12-28 | 2024-07-04 | 中外製薬株式会社 | 環状ペプチドの精製法及び製造法 |
| US12071396B2 (en) | 2019-03-15 | 2024-08-27 | Chugai Seiyaku Kabushiki Kaisha | Method for preparing aromatic amino acid derivative |
| US12163134B2 (en) | 2015-03-13 | 2024-12-10 | Chugai Seiyaku Kabushiki Kaisha | Modified aminoacyl-tRNA synthetase and use thereof |
| WO2025070753A1 (ja) | 2023-09-28 | 2025-04-03 | 中外製薬株式会社 | 混合酸無水物法を用いたペプチド化合物の製造方法 |
| WO2025100405A1 (ja) | 2023-11-06 | 2025-05-15 | 中外製薬株式会社 | 細胞膜透過性と代謝安定性を有する大環状ペプチド化合物、及びこれを含むライブラリ |
| US12312379B2 (en) | 2021-05-07 | 2025-05-27 | Chugai Seiyaku Kabushiki Kaisha | Methods for producing cyclic compounds comprising N-substituted amino acid residues |
| US12312297B2 (en) | 2018-11-07 | 2025-05-27 | Chugai Seiyaku Kabushiki Kaisha | O-substituted serine derivative production method |
| WO2025143082A1 (ja) | 2023-12-27 | 2025-07-03 | 中外製薬株式会社 | 2種以上の添加剤を用いたペプチド化合物の製造方法 |
| WO2025143077A1 (ja) | 2023-12-27 | 2025-07-03 | 中外製薬株式会社 | スタンドアローン型縮合剤を用いたペプチド化合物の製造方法 |
| US12371454B2 (en) | 2019-11-07 | 2025-07-29 | Chugai Seiyaku Kabushiki Kaisha | Cyclic peptide compound having Kras inhibitory action |
| US12391971B2 (en) | 2017-01-31 | 2025-08-19 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptides in cell-free translation system |
| US12404299B2 (en) | 2019-11-07 | 2025-09-02 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide compound comprising highly sterically hindered amino acid |
| US12410212B2 (en) | 2022-05-06 | 2025-09-09 | Chugai Seiyaku Kabushiki Kaisha | Cyclic compound having selective KRAS inhibitory effect on HRAS and NRAS |
| US12637494B2 (en) | 2019-12-12 | 2026-05-26 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide containing non-natural amino acid |
Families Citing this family (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN113929649B (zh) * | 2021-11-22 | 2022-11-01 | 河北化工医药职业技术学院 | 一种科里内酯衍生物的制备方法 |
| CN116924946B (zh) * | 2023-09-15 | 2023-11-21 | 成都泰和伟业生物科技有限公司 | 一种化合物及其制备方法和作为氨基保护试剂的用途 |
Citations (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2013100132A1 (ja) | 2011-12-28 | 2013-07-04 | 中外製薬株式会社 | ペプチド化合物の環化方法 |
| WO2015179823A2 (en) | 2014-05-23 | 2015-11-26 | The California Institute For Biomedical Research | Lung localized inhibitors of alpha(v)beta 6 |
| WO2016148044A1 (ja) | 2015-03-13 | 2016-09-22 | 中外製薬株式会社 | 改変アミノアシルtRNA合成酵素およびその用途 |
| WO2018124162A1 (ja) | 2016-12-28 | 2018-07-05 | 中外製薬株式会社 | 化合物の膜透過性を改善するための自己乳化型製剤 |
| WO2018143145A1 (ja) * | 2017-01-31 | 2018-08-09 | 中外製薬株式会社 | 無細胞翻訳系におけるペプチドの合成方法 |
| WO2018225864A1 (ja) * | 2017-06-09 | 2018-12-13 | 中外製薬株式会社 | 膜透過性の高い環状ペプチド化合物、及びこれを含むライブラリ |
Family Cites Families (10)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| GB8916901D0 (en) * | 1989-07-24 | 1989-09-06 | Sandoz Ltd | Improvements in or relating to organic compounds |
| GB9203886D0 (en) * | 1992-02-24 | 1992-04-08 | Sandoz Ltd | Improvements in or relating to organic compounds |
| US5800804A (en) * | 1993-09-09 | 1998-09-01 | The Gillette Company | O-acyl serines and threonines as deodorants |
| US5672584A (en) * | 1995-04-25 | 1997-09-30 | The University Of Kansas | Cyclic prodrugs of peptides and peptide nucleic acids having improved metabolic stability and cell membrane permeability |
| AU2005265164A1 (en) * | 2004-06-17 | 2006-01-26 | Musc Foundation For Research Development | Non-natural amino acids |
| ES2634431T3 (es) * | 2009-03-16 | 2017-09-27 | The Governing Council Of The University Of Toronto | Moléculas de aminoácidos cíclicos y procedimientos de preparación de las mismas |
| JP5818237B2 (ja) | 2010-09-09 | 2015-11-18 | 国立大学法人 東京大学 | N−メチルアミノ酸およびその他の特殊アミノ酸を含む特殊ペプチド化合物ライブラリーの翻訳構築と活性種探索法 |
| EP2647721B1 (en) * | 2010-12-03 | 2019-02-27 | The University of Tokyo | Peptide with safer secondary structure, peptide library, and production methods for same |
| AU2017267560B2 (en) * | 2016-05-17 | 2022-03-24 | S&T Global Inc. | Novel cyclosporin derivatives and uses thereof |
| US20200392075A1 (en) * | 2018-03-07 | 2020-12-17 | Temple University - Of The Commonwealth System Of Higher Education | Compositions and methods for treatment of insulin resistance |
-
2019
- 2019-12-12 WO PCT/JP2019/048720 patent/WO2020122182A1/ja not_active Ceased
- 2019-12-12 JP JP2020559315A patent/JP7531401B2/ja active Active
- 2019-12-12 EP EP19896830.7A patent/EP3896056A4/en active Pending
- 2019-12-12 US US17/312,296 patent/US20230096766A1/en active Pending
- 2019-12-12 KR KR1020217021220A patent/KR20210102326A/ko active Pending
- 2019-12-12 CN CN201980091739.9A patent/CN113454058B/zh active Active
- 2019-12-12 CN CN202410282724.3A patent/CN118184532A/zh active Pending
-
2024
- 2024-06-27 JP JP2024103888A patent/JP7757469B2/ja active Active
-
2025
- 2025-10-08 JP JP2025170035A patent/JP2025188158A/ja active Pending
Patent Citations (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2013100132A1 (ja) | 2011-12-28 | 2013-07-04 | 中外製薬株式会社 | ペプチド化合物の環化方法 |
| WO2015179823A2 (en) | 2014-05-23 | 2015-11-26 | The California Institute For Biomedical Research | Lung localized inhibitors of alpha(v)beta 6 |
| WO2016148044A1 (ja) | 2015-03-13 | 2016-09-22 | 中外製薬株式会社 | 改変アミノアシルtRNA合成酵素およびその用途 |
| WO2018124162A1 (ja) | 2016-12-28 | 2018-07-05 | 中外製薬株式会社 | 化合物の膜透過性を改善するための自己乳化型製剤 |
| WO2018143145A1 (ja) * | 2017-01-31 | 2018-08-09 | 中外製薬株式会社 | 無細胞翻訳系におけるペプチドの合成方法 |
| WO2018225864A1 (ja) * | 2017-06-09 | 2018-12-13 | 中外製薬株式会社 | 膜透過性の高い環状ペプチド化合物、及びこれを含むライブラリ |
Non-Patent Citations (42)
| Title |
|---|
| "Experimental Chemistry Lecture", vol. 5, MARUZEN CO., article "Thermal Measurements and Equilibrium", pages: 460 |
| "Solid-phase Synthesis Handbook", 1 May 2002, MERCK |
| ALEXANDER ALEX, DAVID S. MILLAN, MANUEL PEREZ, FLORIAN WAKENHUT, GAVIN A. WHITLOCK: "Intramolecular hydrogen bonding to improve membrane permeability and absorption in beyond rule of five chemical space", MEDCHEMCOMM, ROYAL SOCIETY OF CHEMISTRY, UNITED KINGDOM, vol. 2, no. 7, 1 January 2011 (2011-01-01), United Kingdom, pages 669, XP055720117, ISSN: 2040-2503, DOI: 10.1039/c1md00093d * |
| ANDERSON, JC.SCHULTZ, PG.: "Adaptation of an orthogonal archaeal leucyl-tRNA and synthetase pair for four-base, amber, and opal suppression", BIOCHEMISTRY, vol. 42, 2003, pages 9598 - 608, XP002479161, DOI: 10.1021/bi034550w |
| ANGEW. CHEM., vol. 57, 2018, pages 14414 - 14438 |
| BEILSTEIN, JOURNAL OF ORGANIC CHEMISTRY, vol. 13, 2017, pages 2428 - 2441 |
| BERND KUHN, PETER MOHR, MARTIN STAHL: "Intramolecular Hydrogen Bonding in Medicinal Chemistry", JOURNAL OF MEDICINAL CHEMISTRY, AMERICAN CHEMICAL SOCIETY, US, vol. 53, no. 6, 25 March 2010 (2010-03-25), US, pages 2601 - 2611, XP055720106, ISSN: 0022-2623, DOI: 10.1021/jm100087s * |
| CAS , no. 6290-03-5 |
| CAS, no. 741288-31-3 |
| CHIN, JW.CROPP, TA.ANDERSON, JC.MUKHERJI, M.ZHANG, Z.SCHULTZ, PG.CHIN, JW.: "An expanded eukaryotic genetic code", SCIENCE, vol. 301, 2003, pages 964 - 7 |
| DEDKOVA LM ET AL.: "Construction of modified ribosomes for incorporation of D-amino acids into proteins", BIOCHEMISTRY, vol. 45, 2006, pages 15541 - 51 |
| DOI Y ET AL.: "Elongation factor Tu mutants expand amino acid tolerance of protein biosynthesis system", JAM CHEM SOC, vol. 129, 2007, pages 14458 - 62 |
| FUTURE MED. CHEM., vol. 1, 2009, pages 289 - 1310 |
| HARTMAN, MCJOSEPHSON, K.SZOSTAK, JW.: "Enzymatic aminoacylation of tRNA with unnatural amino acids", PROC. NATL. ACAD. SCI. USA., vol. 103, 2006, pages 4356 - 61, XP055311246, DOI: 10.1073/pnas.0509219103 |
| HASHIMOTO, N., NINOMIYA, K., ENDO, T., SISIDO, M.: "Simple and quick chemical aminoacylation of tRNA in cationic micellar solution under ultrasonic agitation", CHEM. COMMUN. (CAMB, no. 34, 2005, pages 4321 - 3 |
| HECKLER, TG.CHANG, LH.ZAMA, Y.NAKA, T.CHORGHADE, MS.HECHT, SM.: "T4 RNA ligase mediated preparation of novel ''chemically misacylated'' tRNAPheS", BIOCHEMISTRY, vol. 23, 1984, pages 1468 - 73 |
| J. AM. CHEM. SOC., vol. 121, 1999, pages 6519 |
| JOSEPHSON, K.HARTMAN, MC.SZOSTAK, JW.: "Ribosomal synthesis of unnatural peptides", J. AM. CHEM. SOC., vol. 127, 2005, pages 11727 - 35, XP055236675, DOI: 10.1021/ja0515809 |
| JOURNAL OF MEDICINAL CHEMISTRY, vol. 34, no. 4, 1991, pages 1297 - 1301 |
| JOURNAL OF ORGANIC CHEMISTRY, vol. 75, 2010, pages 245 |
| JOURNAL OF ORGANIC CHEMISTRY, vol. 79, no. 20, 2014, pages 9699 - 9703 |
| KAWAKAMI, T. ET AL.: "Diverse backbone-cyclized peptides via codon reprogramming", NAT. CHEM. BIOL., vol. 5, 2009, pages 888 - 90, XP055143411, DOI: 10.1038/nchembio.259 |
| KAWAKAMI, T. ET AL.: "Ribosomal synthesis of polypeptoids and peptoid-peptide hybrids", J. AM. CHEM. SOC., vol. 130, 2008, pages 16861 - 3 |
| KIGA, D.SAKAMOTO, K.KODAMA, K.KIGAWA, T.MATSUDA, T.YABUKI, T.SHIROUZU, M.HARADA, YNAKAYAMA, H.TAKIO, K.: "An engineered Escherichia coli tyrosyl-tRNA synthetase for site-specific incorporation of an unnatural amino acid into proteins in eukaryotic translation and its application in a wheat germ cell-free system", PROC. NATL. ACAD. SCI. USA., vol. 99, 2002, pages 9715 - 20 |
| KWON, I ET AL.: "Breaking the degeneracy of the genetic code", J. AM. CHEM. SOC., vol. 125, 2003, pages 7512 - 3 |
| MATTHEAKIS LCBHATT RRDOWER WJ: "An in vitro polysome display system for identifying ligands from very large peptide libraries", PROC NATL ACAD SCI USA., vol. 91, 1994, pages 9022 - 6, XP002043737, DOI: 10.1073/pnas.91.19.9022 |
| MURAKAMI, H.BONZAGNI, NJ.SUGA, H.: "Aminoacyl-tRNA synthesis by a resin-immobilized ribozyme", J. AM. CHEM. SOC., vol. 124, 2002, pages 6834 - 5 |
| MURAKAMI, H.KOUROUKLIS, D.SUGA, H.: "Using a solid-phase ribozyme aminoacylation system to reprogram the genetic code", CHEM. BIOL., vol. 10, 2003, pages 1077 - 84, XP003013895, DOI: 10.1016/j.chembiol.2003.10.010 |
| NEMOTO NMIYAMOTO-SATO EHUSIMI Y: "In vitro virus: bonding of mRNA bearing puromycin at the 3'-terminal end to the C-terminal end of its encoded protein on the ribosome in vitro", FEBS LETT, vol. 414, 1997, pages 405 - 8, XP004366358, DOI: 10.1016/S0014-5793(97)01026-0 |
| NINOMIYA, K.MINOHATA, T.NISHIMURA, M.SISIDO, M.: "In situ chemical aminoacylation with amino acid thioesters linked to a peptide nucleic acid", J. AM. CHEM. SOC., vol. 126, 2004, pages 15984 - 9, XP002337183, DOI: 10.1021/ja048200o |
| ODEGRIP RCOOMBER DELDRIDGE BHEDERERRKUHLMAN PAULLMAN CFITZGERALD KMCGREGOR D: "CIS display: In vitro selection of peptides from librariesof protein-DNA complexes", PROC NATL ACAD SCI USA., vol. 101, 2004, pages 2806 - 10, XP002364430, DOI: 10.1073/pnas.0400219101 |
| PARK HS ET AL.: "Expanding the genetic code of Escherichia coli with phosphoserine", SCIENCE, vol. 333, 2011, pages 1151 - 4, XP055014139, DOI: 10.1126/science.1207203 |
| RAFI, SALMA B. ET AL.: "Predicting and improving the Membrane Permeability of Peptidic Small Molecules", JOURNAL OF MEDICINAL CHEMISTRY, vol. 55, 2012, pages 3163 - 3169, XP055498323, DOI: 10.1021/jm201634q * |
| REIERSEN HLOBERSLI ILOSET GAHVATTUM ESIMONSEN BSTACY JEMCGREGOR DFITZGERALD KWELSCHOF MBREKKE OH: "Covalent antibody display--an in vitro antibody-DNA library selection system", NUCLEIC ACIDSRES., vol. 33, 2005, pages e10, XP055154764, DOI: 10.1093/nar/gni010 |
| ROBERTS RWSZOSTAK JW: "RNA-peptide fusions for the in vitro selection of peptides and proteins", PROC NATL ACAD SCI USA., vol. 94, 1997, pages 12297 - 302, XP002913434, DOI: 10.1073/pnas.94.23.12297 |
| See also references of EP3896056A4 |
| SHIMIZU, Y, INOUE, A., TOMARI, Y, SUZUKI, T., YOKOGAWA, T., NISHIKAWA, K., UEDA, T.: "Cell-free translation reconstituted with purified components", NAT. BIOTECHNOL., vol. 19, 2001, pages 751 - 5, XP037163597, DOI: 10.1038/90802 |
| SHIMIZU, YUEDA, T.: "PURE technology", METHODS MOL. BIOL., vol. 607, 2010, pages 11 - 21 |
| SUBTELNY, AO.HARTMAN, MC.SZOSTAK, JW.: "Ribosomal synthesis of N-methyl peptides", J. AM. CHEM. SOC., vol. 130, 2008, pages 6131 - 6, XP055376645, DOI: 10.1021/ja710016v |
| TAWFIK DSGRIFFITHS AD: "Man-made cell-like compartments for molecular evolution", NAT BIOTECHNOL., vol. 16, 1998, pages 652 - 6, XP002669764, DOI: 10.1038/nbt0798-652 |
| THE PRACTICE OF MEDICINAL CHEMISTRY, vol. 1, pages 87 |
| YAMAGUCHI J, NAIMUDDIN M, BIYANI M, SASAKI T, MACHIDA M, KUBO T, FUNATSU T, HUSIMI Y, NEMOTO N.: "cDNA display: a novel screening method for functional disulfide-rich peptides by solid-phase synthesis and stabilization of mRNA-protein fusions", NUCLEIC ACIDS RES., vol. 37, no. 16, 2009, pages e108, XP008154704, DOI: 10.1093/nar/gkp514 |
Cited By (36)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US11891457B2 (en) | 2011-12-28 | 2024-02-06 | Chugai Seiyaku Kabushiki Kaisha | Peptide-compound cyclization method |
| US12415835B2 (en) | 2011-12-28 | 2025-09-16 | Chugai Seiyaku Kabushiki Kaisha | Peptide-compound cyclization method |
| US12163134B2 (en) | 2015-03-13 | 2024-12-10 | Chugai Seiyaku Kabushiki Kaisha | Modified aminoacyl-tRNA synthetase and use thereof |
| US12391971B2 (en) | 2017-01-31 | 2025-08-19 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptides in cell-free translation system |
| US11787836B2 (en) | 2017-06-09 | 2023-10-17 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptide containing N-substituted amino acid |
| US11542299B2 (en) | 2017-06-09 | 2023-01-03 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptide containing N-substituted amino acid |
| US12281141B2 (en) | 2017-06-09 | 2025-04-22 | Chugai Seiyaku Kabushiki Kaisha | Method for synthesizing peptide containing N-substituted amino acid |
| US11492369B2 (en) | 2017-12-15 | 2022-11-08 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide, and method for processing bases |
| US12312297B2 (en) | 2018-11-07 | 2025-05-27 | Chugai Seiyaku Kabushiki Kaisha | O-substituted serine derivative production method |
| US12331072B2 (en) | 2018-11-30 | 2025-06-17 | Chugai Seiyaku Kabushiki Kaisha | Deprotection method and resin removal method in solid-phase reaction for peptide compound or amide compound, and method for producing peptide compound |
| US11732002B2 (en) | 2018-11-30 | 2023-08-22 | Chugai Seiyaku Kabushiki Kaisha | Deprotection method and resin removal method in solid-phase reaction for peptide compound or amide compound, and method for producing peptide compound |
| US12071396B2 (en) | 2019-03-15 | 2024-08-27 | Chugai Seiyaku Kabushiki Kaisha | Method for preparing aromatic amino acid derivative |
| US12371454B2 (en) | 2019-11-07 | 2025-07-29 | Chugai Seiyaku Kabushiki Kaisha | Cyclic peptide compound having Kras inhibitory action |
| US12404299B2 (en) | 2019-11-07 | 2025-09-02 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide compound comprising highly sterically hindered amino acid |
| US12637494B2 (en) | 2019-12-12 | 2026-05-26 | Chugai Seiyaku Kabushiki Kaisha | Method for producing peptide containing non-natural amino acid |
| KR20230124935A (ko) | 2020-12-25 | 2023-08-28 | 추가이 세이야쿠 가부시키가이샤 | N-치환-아미노산 잔기를 포함하는 펩타이드 화합물의제조 방법 |
| WO2022138891A1 (ja) | 2020-12-25 | 2022-06-30 | 中外製薬株式会社 | N-置換-アミノ酸残基を含むペプチド化合物の製造方法 |
| WO2022145444A1 (ja) | 2020-12-28 | 2022-07-07 | 中外製薬株式会社 | アミノ酸の固相合成用樹脂への担持方法 |
| KR20230125807A (ko) | 2020-12-28 | 2023-08-29 | 추가이 세이야쿠 가부시키가이샤 | 아미노산의 고상 합성용 수지에의 담지 방법 |
| JPWO2022234853A1 (ja) * | 2021-05-07 | 2022-11-10 | ||
| WO2022234853A1 (ja) * | 2021-05-07 | 2022-11-10 | 中外製薬株式会社 | Hrasおよびnrasに対して選択的なkras阻害作用を有する環状化合物 |
| WO2022234850A1 (ja) | 2021-05-07 | 2022-11-10 | 中外製薬株式会社 | 環状ペプチド化合物を含む製剤及びその製造方法 |
| KR20230169093A (ko) | 2021-05-07 | 2023-12-15 | 추가이 세이야쿠 가부시키가이샤 | 환상 펩타이드 화합물을 포함하는 제제 및 그의 제조 방법 |
| KR20230020548A (ko) | 2021-05-07 | 2023-02-10 | 추가이 세이야쿠 가부시키가이샤 | 환상 펩타이드 화합물을 포함하는 제제 및 그의 제조 방법 |
| US12312379B2 (en) | 2021-05-07 | 2025-05-27 | Chugai Seiyaku Kabushiki Kaisha | Methods for producing cyclic compounds comprising N-substituted amino acid residues |
| KR20240113971A (ko) | 2021-12-28 | 2024-07-23 | 추가이 세이야쿠 가부시키가이샤 | N-알킬 아미노산, 및 n-알킬 아미노산을 포함하는 펩타이드의 제조 방법 |
| WO2023127869A1 (ja) | 2021-12-28 | 2023-07-06 | 中外製薬株式会社 | N-アルキルアミノ酸、およびn-アルキルアミノ酸を含むペプチドの製造方法 |
| US12410212B2 (en) | 2022-05-06 | 2025-09-09 | Chugai Seiyaku Kabushiki Kaisha | Cyclic compound having selective KRAS inhibitory effect on HRAS and NRAS |
| WO2024043251A1 (ja) | 2022-08-23 | 2024-02-29 | 富士フイルム株式会社 | 環状ペプチドの細胞膜透過性の予測方法 |
| WO2024096023A1 (ja) | 2022-11-01 | 2024-05-10 | 中外製薬株式会社 | ジベンゾフルベンまたはジベンゾフルベン誘導体の除去方法 |
| WO2024143514A1 (ja) | 2022-12-28 | 2024-07-04 | 中外製薬株式会社 | 環状ペプチドの精製法及び製造法 |
| EP4631955A1 (en) | 2022-12-28 | 2025-10-15 | Chugai Seiyaku Kabushiki Kaisha | Purification method and production method of cyclic peptide |
| WO2025070753A1 (ja) | 2023-09-28 | 2025-04-03 | 中外製薬株式会社 | 混合酸無水物法を用いたペプチド化合物の製造方法 |
| WO2025100405A1 (ja) | 2023-11-06 | 2025-05-15 | 中外製薬株式会社 | 細胞膜透過性と代謝安定性を有する大環状ペプチド化合物、及びこれを含むライブラリ |
| WO2025143082A1 (ja) | 2023-12-27 | 2025-07-03 | 中外製薬株式会社 | 2種以上の添加剤を用いたペプチド化合物の製造方法 |
| WO2025143077A1 (ja) | 2023-12-27 | 2025-07-03 | 中外製薬株式会社 | スタンドアローン型縮合剤を用いたペプチド化合物の製造方法 |
Also Published As
| Publication number | Publication date |
|---|---|
| JP2025188158A (ja) | 2025-12-25 |
| KR20210102326A (ko) | 2021-08-19 |
| JPWO2020122182A1 (ja) | 2021-10-28 |
| EP3896056A1 (en) | 2021-10-20 |
| EP3896056A4 (en) | 2023-03-15 |
| JP7531401B2 (ja) | 2024-08-09 |
| JP2024124450A (ja) | 2024-09-12 |
| US20230096766A1 (en) | 2023-03-30 |
| CN113454058B (zh) | 2024-03-22 |
| CN113454058A (zh) | 2021-09-28 |
| JP7757469B2 (ja) | 2025-10-21 |
| CN118184532A (zh) | 2024-06-14 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| JP7757469B2 (ja) | 分子内水素結合可能な官能基を有するアミノ酸とそれらのアミノ酸を含むペプチド化合物、およびそれらの製造方法 | |
| JP7611279B2 (ja) | 膜透過性の高い環状ペプチド化合物、及びこれを含むライブラリ | |
| JP7526243B2 (ja) | 無細胞翻訳系におけるペプチドの合成方法 | |
| JP7351968B2 (ja) | チオール基をアミノ基近傍に有するアミノ酸をn末端に持つ非環状ペプチド-核酸複合体、そのライブラリー、およびそれから誘導される環状ペプチド-核酸複合体ライブラリーの製造方法 | |
| TW201900670A (zh) | 胜肽化合物之環化方法 | |
| Kawamura et al. | Design and synthesis of the stabilized analogs of belactosin A with the unnatural cis-cyclopropane structure | |
| Jacobsen et al. | Synthesis of cyclic peptide analogues of the 310 helical Pro138-Gly144 segment of human aquaporin-4 by olefin metathesis | |
| US20240409524A1 (en) | Compound having photocleavable moiety, conjugate, and screening method using same | |
| HK40104843A (en) | Amino acid having functional group capable of intramolecular hydrogen bonding, peptide compound containing same and method for production thereof | |
| Zhang et al. | Synthesis of Enediamino amino acid analogues and peptides | |
| WO2025100405A1 (ja) | 細胞膜透過性と代謝安定性を有する大環状ペプチド化合物、及びこれを含むライブラリ | |
| Bellucci et al. | Synthesis of hexafluorovaline-containing di-and tripeptides | |
| Elbatrawi | Total Synthesis and Conformational Analysis of Cyclic Hydrazino Acid Natural Products and Peptidomimetics | |
| Robertson | Chemical biology tools to explore the activities and mechanisms of Class I HDAC corepressor complexes | |
| Gardiner | Conformationally restricted amino acid analogues based on lactams | |
| Marimganti | Synthesis and conformational analysis of jasplakinolide analogues and approach towards the synthesis of the stereotetrad of cruentaren A | |
| US20030073721A1 (en) | Peptide beta-strand mimics based on 1,2-dihydro-3(6H)-pyridinone |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 19896830 Country of ref document: EP Kind code of ref document: A1 |
|
| ENP | Entry into the national phase |
Ref document number: 2020559315 Country of ref document: JP Kind code of ref document: A |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| ENP | Entry into the national phase |
Ref document number: 20217021220 Country of ref document: KR Kind code of ref document: A |
|
| ENP | Entry into the national phase |
Ref document number: 2019896830 Country of ref document: EP Effective date: 20210712 |