EP3752482B1 - Verfahren zur herstellung von bis-cholin-tetrathiomolybdat - Google Patents
Verfahren zur herstellung von bis-cholin-tetrathiomolybdat Download PDFInfo
- Publication number
- EP3752482B1 EP3752482B1 EP19706505.5A EP19706505A EP3752482B1 EP 3752482 B1 EP3752482 B1 EP 3752482B1 EP 19706505 A EP19706505 A EP 19706505A EP 3752482 B1 EP3752482 B1 EP 3752482B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- compound
- reaction mixture
- process according
- iii
- reaction
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Active
Links
Images


Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
- C07F11/00—Compounds containing elements of Groups 6 or 16 of the Periodic Table
- C07F11/005—Compounds containing elements of Groups 6 or 16 of the Periodic Table compounds without a metal-carbon linkage
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C215/00—Compounds containing amino and hydroxy groups bound to the same carbon skeleton
- C07C215/02—Compounds containing amino and hydroxy groups bound to the same carbon skeleton having hydroxy groups and amino groups bound to acyclic carbon atoms of the same carbon skeleton
- C07C215/40—Compounds containing amino and hydroxy groups bound to the same carbon skeleton having hydroxy groups and amino groups bound to acyclic carbon atoms of the same carbon skeleton with quaternised nitrogen atoms bound to carbon atoms of the carbon skeleton
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C209/00—Preparation of compounds containing amino groups bound to a carbon skeleton
- C07C209/68—Preparation of compounds containing amino groups bound to a carbon skeleton from amines, by reactions not involving amino groups, e.g. reduction of unsaturated amines, aromatisation, or substitution of the carbon skeleton
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C213/00—Preparation of compounds containing amino and hydroxy, amino and etherified hydroxy or amino and esterified hydroxy groups bound to the same carbon skeleton
- C07C213/08—Preparation of compounds containing amino and hydroxy, amino and etherified hydroxy or amino and esterified hydroxy groups bound to the same carbon skeleton by reactions not involving the formation of amino groups, hydroxy groups or etherified or esterified hydroxy groups
Definitions
- the present disclosure relates to an improved process for preparing bis-choline tetrathiomolybdate (BC-TTM).
- Bis-choline tetrathiomolybdate can act as protein aggregator with an affinity for copper, and is thus considered potentially useful in the treatment of various diseases associated with aberrant vascularization, copper metabolism disorders, neurodegenerative disorders, obesity or NF- ⁇ B dysregulation.
- One area of interest is the use of bis-choline tetrathiomolybdate in the treatment of Wilson disease.
- Wilson disease is an autosomal recessive genetic disorder in which copper accumulates in tissues; this manifests as neurological or psychiatric symptoms and liver disease. It is treated by medication that reduces copper absorption or removes the excess copper from the body.
- U.S. Patent Application Publication No. US 2004/0019087 discloses various thiomolybdate derivatives and their use in pharmaceutical compositions for the treatment or prevention of diseases associated with aberrant vascularization, copper metabolism disorders, neurodegenerative disorders, obesity or NF- ⁇ B dysregulation, and more particularly Wilson disease.
- ClinicalTrials.gov NCT02273596.
- the ' 865 Patent further discloses that the product was precipitated from solution by the addition of isopropanol, and then washed with isopropanol, ethanol and diethyl ether.
- bis-choline tetrathiomolybdate was prepared from choline hydroxide and ammonium tetrathiomolybdate according to that method illustrated in Scheme 1 at scales ranging from 2 g to 140 g, with yields ranging from 65% to 83%.
- the ' 865 Patent also discloses an alternative method for the synthesis of bis-choline tetrathiomolybdate in which two equivalents of choline hydroxide are added to one equivalent of ammonium molybdate in an aqueous solution, and then hydrogen sulfide gas (H 2 S) is bubbled through the solution at room temperature (see '865 Patent, Examples 6.22 and 6.23). It is disclosed that, after purging the solution with nitrogen gas, the solvent (water) was removed repeatedly under reduced pressure to remove ammonia, and that the product needed to be redissolved in water and filtered to remove insoluble molybdenum sulfide. The '865 Patent disclosed that the product was recrystallized from water and isopropanol, and washed with ethanol and diethyl ether, to provide a 90% yield on a 1.5 g laboratory bench scale.
- the present invention provides a process for the production of compound (I): wherein the process comprises:
- the process of the disclosure provides the reaction product in good yield and with good purity in an economical reaction time without the need to use hydrogen disulphide.
- X -1 is OH -1 .
- the process further comprises monitoring the evolution of ammonia during the step of subjecting the first reaction mixture to a reduced pressure.
- monitoring the evolution of ammonia comprises measuring the pH of vapour of a gas phase in contact with the first reaction mixture.
- achievement of pH of about 7 indicates that the second reaction period is complete.
- isolating compound (I) comprises adding from 2 to 4 volumes of ethanol to the second reaction mixture.
- the molar ratio of compound (II) to compound (III) is in the range of 4 to 20, 5 to 20, or 5 to 15.
- the total reaction time of the process is reduced by at least 50% compared to a process in which compound (II) is not used in a molar excess.
- compound (III) does not significantly decompose during the process.
- the first mixture of compound (III) in water is a slurry of compound (III) in water.
- the step of combining compound (II) with a first mixture of compound (II) in water is carried out at a temperature of 10°C or less, preferably at a temperature of from 0°C to 10°C.
- compound (II) is combined with the first mixture of compound (III) as an aqueous solution.
- the step of subjecting the first reaction mixture to a reduced pressure is carried out at a temperature of 35 °C or less, preferably at a temperature of from 5 °C to 35 °C.
- the step of subjecting the first reaction mixture to a reduced pressure comprises subjecting the first reaction mixture to a pressure of 100 mbar or less, preferably to a pressure of from 0.1 mbar to 100 mbar.
- the step of isolating compound (I) from the second reaction mixture comprises precipitating compound (I) from the second reaction mixture and optionally filtering the second reaction mixture to obtain compound (I) as a solid.
- the process further comprises a recrystallization step, followed by optional polish filtering, optional washing with ethanol, drying of the resultant solid, and measurement of resulting particle size of the resultant solid.
- the resultant solid has a 50th percentile of particle size (D 50 ) of at least 30 ⁇ m, at least 40 ⁇ m, between 30 ⁇ m and 90 ⁇ m, or between 30 ⁇ m and 70 ⁇ m.
- the resultant solid has a 90th percentile of particle size (D 90 ) of between 60 ⁇ m and 550 ⁇ m, between 400 ⁇ m and 550 ⁇ m, between 75 ⁇ m and 135 ⁇ m, or between 75 ⁇ m and 100 ⁇ m.
- the resultant solid has a D 50 of less than 35 ⁇ m.
- the resultant solid has a D 90 of less than 75 ⁇ m.
- the resultant solid has a D 90 -to-D 50 ratio less than 3.0, optionally less than or equal to 2.5 or optionally less than 2.0.
- the disclosure further provides a pharmaceutical composition comprising at least one compound prepared according to any of the preceding claims and at least one pharmaceutically acceptable excipient.
- the disclosure further provides a method of treating a patient with at least one disease associated with aberrant vascularization, copper metabolism disorders, neurodegenerative disorders, obesity, and NF- ⁇ B dysregulation, comprising administering a therapeutically effective amount of a compound prepared according to any of the preceding claims.
- the copper metabolism disorder may be Wilson disease.
- the disclosure further provides a kit for treating a patient with Wilson disease, the kit comprising a compound prepared according to any of the preceding claims, at least one pharmaceutically acceptable excipient, and instructions for administration to the patient.
- the present disclosure provides an improved process for the large-scale synthesis of bis-choline tetrathiomolybdate, comprising treating an aqueous mixture of an ammonium thiomolybdate with an excess of a choline salt, exposing the resulting reaction mixture to a reduced pressure, and isolating the bis-choline tetrathiomolybdate from the resulting reaction mixture, the step of isolating the bis-choline tetrathiomolybdate comprising adding ethanol to the reaction mixture.
- the choline salt is choline hydroxide (i.e. the X -1 counter-ion is OH -1 ).
- the choline salt is [N(Me) 3 (CH 2 CH 2 OH)][OH].
- the word “a,” “an” or “plurality” before a noun represents one or more of the particular noun.
- the phrase “a compound” represents “one or more compounds.” Where the plural form (e.g. compounds, salts) is used, this includes the singular (e.g. a single compound, a single salt). "A compound” does not exclude that (e.g. in a pharmaceutical formulation) where more than one compound or salt is present.
- the term "isolating” refers to any means of separating a compound (e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate) from a solution comprising a solvent and, in some cases, impurities. Isolating may include precipitating the compound from the solution (e.g. with ethanol or an ethanol/water mixture) and/or filtering the resulting solid compound. Isolating may include one or more washing steps. Isolating removes at least about 50%, 60%, 70%, 80%, 82%, 85%, 87%, 90%, 92%, 95%, 96%, 97%, 98%, 99%, 99.5%, or 99.9% of the solvent and/or impurities in the solution.
- a compound e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate
- Isolating may include precipitating the compound from the solution (e.g. with ethanol or an ethanol/water mixture) and/
- patient and “patients” are understood to represent any subject (especially a mammal, especially a human) for administration of a compound according to the disclosure, unless explicitly stated otherwise.
- precipitating means to cause a compound (e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate) to separate from a solution into solid form.
- a compound may be precipitating from aqueous solution by, for example, adding ethanol or a water/ethanol mixture to the solution. In some cases, precipitating may cause one compound to separate from solution while a second compound (e.g. an impurity) remains in solution.
- Precipitating may cause at least about 50%, 60%, 70%, 80%, 82%, 85%, 87%, 90%, 92%, 95%, 96%, 97%, 98%, 99%, 99.5%, or 99.9% of the compound to separate from the solution as a solid (e.g. a crystalline solid).
- a solid e.g. a crystalline solid
- the term "significantly” refers to a change that is statistically significant, not due to chance alone, and observable using chemical or physical methods.
- a compound e.g., (NH 4 ) 2 MoS 4
- does not significantly decompose means that less than about 25%, 20%, 15%, 10%, 7%, 5%, 4%, 3%, 1%, or 0.5% w/w of the compound decomposes.
- the term "complete" with respect to a chemical reaction means that the chemical reaction has proceeded to completion, or nearly to completion.
- a chemical reaction is said to be complete when allowing additional time to elapse, without changing process parameters (such as temperature) or adding new reagents, will not cause a significant increase in the yield of the product of the chemical reaction.
- Complete may mean that the reaction is 85%, 87%, 90%, 92%, 95%, 96%, 97%, 98%, 99%, or 99.5% complete, i.e. that continuing the chemical reaction longer would cause at most a 15%, 13%, 10%, 8%, 5%, 4%, 3%, 2%, 1%, or 0.5% increase in the yield of the product of the chemical reaction.
- yield refers to the amount of product obtained in a chemical reaction.
- the percentage yield also termed fractional yield or relative yield
- the theoretical yield is the amount of product that would be obtained if all reactants in the chemical reaction converted stoichiometrically to product with no remaining reactants and no reactant lost to side reactions. Yield may also be reported as absolute yield in grams (mass yield) or in moles (molar yield). With respect to a process involving multiple chemical reactions, yield refers to the amount of product obtained for the entire process.
- the percentage yield of a process is the amount of desired product obtained from the process divided by the amount of product that would be obtain if each chemical reaction in the process achieved the theoretical yield for that reaction step.
- high yield refers to a process with a percentage yield of at least 70%, 75%, 80%, 85%, 90%, or 95%.
- large scale refers to a chemical reaction, or process involving multiple chemical reactions, that yields at least one kilogram (kg) of a desired product.
- a large scale process may yield 1 kg, 2.5 kg, 5 kg, 7.5 kg, 10 kg, 12 kg, 15kg, 20 kg, 25 kg or more of the desired product (e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate).
- small scale refers to a chemical reaction, or process involving multiple chemical reactions, that yields less than 100 grams (g) of a desired product.
- a small scale process may yield about 100 g, 50 g, 40 g, 30 g, 20 g, 10 g, or less of the desired product (e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate).
- high level of purity refers to a purity of at least about 95%.
- a compound having high level of purity may be at least about 80%, 82%, 85%, 87%, 90%, 92%, 95%, 96%, 97%, 98%, 99%, 99.5%, or 99.9% free of impurities.
- the level of purity as a percentage may be calculated by dividing the amount of the desired compound (e.g. a thiomolybdate derivative, especially bis-choline tetrathiomolybdate) by the total amount of the desired compound plus any impurities.
- HPLC high-performance liquid chromatography
- the term “Dx(10)/ ⁇ m” refers to the 10th percentile of particle size (D 10 ) measured in micrometers ( ⁇ m).
- the terms “Dx(50)/ ⁇ m” or “median particle size” refer to the 50 th percentile particle size (D 50 ).
- the term “Dx(90)/ ⁇ m” refers to the 90th percentile of particle size (D 90 ).
- Wilson disease As used herein, the terms "Wilson disease,” “Wilson's disease,” or “WD” are equivalent and represent an autosomal recessive genetic disorder in which copper accumulates in tissues of a patient.
- the reduced pressure removes ammonia released during the reaction, and thus drives the reaction to completion following Le C.lier's principle (the ammonium tetrathiomolybdate starting material and the bis-choline tetrathiomolybdate product being in equilibrium). Subjecting the reaction mixture to a reduced pressure is also believed to assist in reducing reaction time, thereby allowing the bis-choline tetrathiomolybdate product to be isolated more quickly. A shorter reaction time helps to avoid decomposition of the thiomolybdate species during the reaction.
- the process of the disclosure works very well and reliably affords the bis-choline tetrathiomolybdate product in both high yield and high purity.
- the combination step comprises combining a molar excess of compound (II) with compound (III), i.e. a molar ratio of compound (II) to compound (III) of Z: 1, wherein Z is greater than 2 (i.e. more than two parts of compound (II) for each one part of compound (III), is used.
- a molar excess of compound (II) with compound (III) i.e. a molar ratio of compound (II) to compound (III) of Z: 1, wherein Z is greater than 2 (i.e. more than two parts of compound (II) for each one part of compound (III)
- Z is greater than 2 (i.e. more than two parts of compound (II) for each one part of compound (III)
- the present inventors found that when the small-scale preparation of bis-choline tetrathiomolybdate was carried out in methanol with a molar ratio of choline hydroxide to ammonium tetrathiomolybdate of 10:1, the time taken for complete release of ammonia was significantly reduced, from 180 minutes to 80 minutes.
- the molar ratio of compound (II) to compound (III) is Z:1, wherein Z is 3 or greater, or 4 or greater, or 5 or greater, or 6 or greater, for example 7 or greater.
- Z is up to 20, especially up to 15, for example up to 12.
- Z may be in the range of from above 2 to 20, for example in the range of 3, or 4, or 5, or 6, to 20, for example 8, or 10, or 12 to 20, for example 3, or 4, or 5, or 6, or 8, or 10 or 12 to 15, for example 12 to 15.
- Z is in the range of from 5 to 20, preferably from 5 to 15, especially from 5 to 12, for example from 5 to 7.
- the total reaction time of the process is reduced by at least 50% compared to a process in which Z is 2 or less.
- the time taken for the second reaction period to be complete is reduced by at least 50% compared to a process in which Z is 2 or less.
- compound (III) does not significantly decompose during the process. It will be understood that the expected reaction of compound (III) with compound (II) to form compound (I) is not decomposition of compound (III). For example, it may be that decomposition of compound (III) is thermal decomposition, such as thermal decomposition into unwanted side products. It will be understood that 'unwanted side products' may be molybdenum species other than compound (I) or compound (III).
- the step of isolating compound (I) from the second reaction mixture comprises adding ethanol to the second reaction mixture.
- adding ethanol to the second reaction mixture aids the isolation of compound (I) in a relatively pure form.
- adding ethanol to the second reaction mixture causes the precipitation of compound (I) while leaving compound (II) in solution.
- separating compound (I) from a reaction mixture comprising compound (I) and compound (II) may be particularly useful when an excess of compound (II) is used in the preparation of compound (I).
- choline hydroxide has good solubility in ethanol-water mixtures, while bis-choline tetrathiomolybdate has poor solubility in ethanol-water mixtures, and that the use of ethanol in the isolating step provides bis-choline tetrathiomolybdate in high yield with a high purity. More particularly, the use of ethanol instead of isopropanol to precipitate bis-choline tetrathiomolybdate is advantageous. This is surprising because if bis-choline tetrathiomolybdate is recrystallized from an alcohol, the degree of purification is comparable whether ethanol or isopropanol is used. However, if isopropanol is used instead of ethanol in the isolating step of the present disclosure, precipitation of excess choline hydroxide occurs, leading to a relatively impure product compared with the product obtained using ethanol.
- the isolating step comprises treating the second mixture with 2 to 4 volumes, for example about 3 volumes, of ethanol to precipitate compound (I).
- 2 to 4 volumes for example about 3 volumes
- ethanol to precipitate compound (I).
- the present inventors have found that the use of 3 volumes of ethanol gave a good yield in the preparation of bis-choline tetrathiomolybdate.
- the compound (I) can be isolated as a solvate, for example a hydrate.
- the step of combining compound (II) with a first mixture of compound (III) in water is carried out at a temperature of 40°C or less, such as 35°C or less, for example at room temperature or less.
- the step of combining compound (II) with a first mixture of compound (III) in water is carried out at a temperature of 10°C or less, such as 8°C or less, for example 5°C or less.
- the step of combining compound (II) with a first mixture of compound (III) in water is carried out at a temperature of from 0°C to 10°C, for example 0°C to 5°C.
- the first reaction period is a period of 90 minutes or less, for example one hour or less.
- the compound (II) is added to the first mixture of compound (III) in water.
- the step of subjecting the first reaction mixture to a reduced pressure is carried out at a temperature of 37°C or less, for example 35°C or less, such as 33°C or less, for example 30°C or less.
- the step of subjecting the first reaction mixture to a reduced pressure is carried out at a temperature of from 5°C to 35°C, for example 10°C to 33°C, for example 25°C to 30°C, or alternatively 10°C to 20°C.
- the second reaction period is a period of from 30 minutes to 10 hours, from 1 to 8 hours, for example 2 to 6 hours.
- the first mixture of compound (III) in water is a slurry of compound (III) in water.
- the present inventors have found that it is not necessary for compound (III) to be fully dissolved in water during the addition step.
- the use of a slurry enables the volume of water to be smaller than would be needed for a solution, and the smaller volume of water assists in control of the temperature. It has been found that use of compound (III) in a slurry does not have any detrimental effects on the rate of reaction, the yield, or the purity of the product.
- compound (II) is added to the first mixture of compound (III) as a solution.
- the solution may be substantially free, or free, from methanol, for example it may be an aqueous solution.
- the present inventors have found that the step of isolating compound (I) from the second reaction mixture can be carried out more easily, and that the process affords a higher yield, when compound (II) is added as a solution substantially free from methanol. Without wishing to be bound by theory, the present inventors believe that the presence of methanol can prevent precipitation of compound (I) from the second reaction mixture during the isolating step.
- an aqueous solution of compound (II) is added dropwise to the aqueous mixture of compound (III). Dropwise addition facilitates control of the temperature of the resulting reaction mixture.
- a reduced pressure is a pressure lower than atmospheric pressure.
- the step of subjecting the first reaction mixture to a reduced pressure comprises subjecting the first reaction mixture to a pressure of 100 mbar or less, such as 50 mbar or less, for example 25 mbar or less.
- the step of subjecting the first reaction mixture to a reduced pressure comprises subjecting the first reaction mixture to a pressure of from 0.1 mbar to 100 mbar, such as from 0.5 mbar to 50 mbar, for example from 1 mbar to 25 mbar.
- the present inventors believe that, since ammonia is formed during the reaction, the reaction of compound (III) with compound (II) can be monitored by following the formation of ammonia.
- the present inventors found that the pH of the vapour formed during the reaction can be estimated by applying a stream of nitrogen gas over the surface of the reaction mixture and testing the N 2 stream using moist pH indicator sticks. In a test reaction, a pH of 10 was measured in the vapour 5 minutes after the reaction started. The first reaction mixture was left to stir overnight at 30°C, after which time the vapour was found to have a pH of 7, indicating that no more ammonia was being released.
- the process comprises monitoring the evolution of ammonia during the step of subjecting the first reaction mixture to a reduced pressure. It may be that monitoring the evolution of ammonia during the step of subjecting the first reaction mixture to a reduced pressure provides an indication on the progress of the reaction, for example allowing the step of isolating compound (III) from the second reaction mixture to be commenced as soon as the reaction is complete. It will be appreciated that it is generally desirable to minimise the period during which a reaction product is maintained in solution.
- the step of monitoring the evolution of ammonia comprises detecting the presence of ammonia in the gas phase, for example at intervals, during the step of subjecting the first reaction mixture to a reduced pressure.
- the step of monitoring the evolution of ammonia comprises determining the basicity of the gas phase, for example by measuring the pH of the gas phase.
- the process comprises subjecting the first reaction mixture to a reduced pressure until substantially no ammonia is detected in the gas phase.
- the process comprises subjecting the first reaction mixture to a reduced pressure until the pH of the gas phase is found to be essentially neutral, such as having a pH of from 6 to 8, for example a pH of about 7.
- the exact time necessary depends on the concentration of the reagents and the temperature of the reaction mixture.
- the second reaction period is a period of from 30 minutes to 10 hours, from 1 to 8 hours, for example 2 to 6 hours.
- achievement and/or detection of a pH of about 7 indicates that the second reaction period is complete.
- the step of isolating compound (I) from the second reaction mixture comprises precipitating compound (I) from the second reaction mixture.
- the step of isolating compound (I) from the second reaction mixture may comprise filtering the second reaction mixture to obtain compound (I) as a solid.
- Compound (I) can be further recrystallized, for example as described in PCT/EP2019/052940 .
- Compound (I) can be recrystallized by (a) providing a solution Compound (I); (b) heating the solution of Compound (I) to a first temperature between about 35 °C and 50 °C; (b) gradually adding ethanol or isopropanol to the solution of Compound (I) over a time period of at least one hour; (c) cooling the solution of Compound (I) to a second temperature between about 10 °C and 25 °C to produce solid Compound (I); (d) filtering the solid Compound (I); (e) drying the solid Compound (I) at a third temperature between about 20 °C and 30 °C under reduced pressure.
- the crystalline particles of Compound (I) have a 50 th percentile of particle size (D 50 ) of at least 30 ⁇ m. In other embodiments, the crystalline particles have a D 50 of at least 40 ⁇ m. In other embodiments, the crystalline particles have a D 50 of between 30 ⁇ m and 90 ⁇ m. In other embodiments, the crystalline particles have a D 50 of between 30 ⁇ m and 70 ⁇ m.
- the crystalline particles of Compound (I) have a 50 th percentile of particle size (D 50 ) of at least 30 ⁇ m, or at least 40 ⁇ m, or between 30 ⁇ m and 70 ⁇ m, and a 90th percentile of particle size (D 90 ) of between 75 ⁇ m and 550 ⁇ m.
- the D 90 is between 400 ⁇ m and 550 ⁇ m.
- the D 90 is between 75 ⁇ m and 135 ⁇ m.
- the D 90 is between 75 ⁇ m and 100 ⁇ m.
- the crystalline particles have a D 90 -to-D 50 ratio of less than 2.5.
- the D 50 is less than 35 ⁇ m.
- the D 90 is less than 75 ⁇ m.
- the crystalline particles have a 10th percentile of particle size (D 10 ) of less than 15 ⁇ m.
- Bis-choline tetrathiomolybdate, compound (I) can exist in a number of different crystal forms.
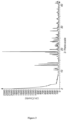
- One crystal form, referred to as Form 1 has an X-ray powder diffraction pattern, as obtained with CuK ⁇ 1-radiation, with characteristic peaks at diffraction angles (°2theta) of 12.64 ⁇ 0.15 and 18.89 ⁇ 0.15; more specifically with characteristic peaks at diffraction angles (°2theta) of 12.64 ⁇ 0.15; 13.84 ⁇ 0.15; 18.64 ⁇ 0.15; 18.89 ⁇ 0.15; and 19.86 ⁇ 0.15; and more specifically having an X-ray powder diffraction pattern essentially as set out in Table 3 in Example 4, or essentially as set out in Figure 3 .
- Form 2 Another crystal form, referred to as Form 2, has a X-ray powder diffraction pattern, as obtained with CuK ⁇ 1-radiation, with characteristic peaks at diffraction angles (°2theta) of 12.84 ⁇ 0.15 and 19.08 ⁇ 0.15; more specifically with characteristic peaks at diffraction angles (°2theta) of 11.00 ⁇ 0.15; 12.84 ⁇ 0.15; 16.02 ⁇ 0.15; 17.28 ⁇ 0.15; and 19.08 ⁇ 0.15; and more specifically having an X-ray powder diffraction pattern essentially as set out in Table 4 in Example 5, or essentially as set out in Figure 2 .
- Form 1 may be prepared by a method comprising:
- the solution in step a) is preferably kept at a temperature of from about 10 to about 35 °C, or from about 10 °C to about 25 °C.
- An alcohol other than ethanol could be used in step a) above.
- the solution in step b) is preferably stirred for about 15 minutes to about 24 hours, for example for about 15 minutes to about 2 hours, for example for about 15 minutes to about 45 minutes.
- the alcohol in step c) is preferably ethanol.
- Alternative solvents or solvent combinations to ethyl acetate include, but are not limited to, at least one selected from heptane, cumene, anisole, butyl acetate, isopropyl acetate, propyl acetate, methyl isobutylketone, methyl ethylketone, isopropyl alcohol, hexane, cyclohexane, 1,4-dioxolane, toluene, tetralin, chlorobenzene, tetrahydrofuran, dichloromethane, acetonitrile, dimethylamine, cyclohexanone, diisopropyl ether, tetrahydrofuran +5%H 2 O, isopropylalcohol+5%H 2 O, acetone+5%H 2 O, 1,2,3,4-tetrahydronaftalen, morpholine, mesitylene, ethyl formate, tetradecafluoro
- Form 2 may for example be prepared by a method comprising:
- step b) The isolation of material in step b) is carried out at 56.8 °C.
- Bis-choline tetrathiomolybdate Form 1 and Form 2 are both crystalline, non-solvated, forms.
- reaction solution was transferred to a 5L 3-necked round bottomed flask through a sintered glass-funnel (Por. 3). No residues were found on the sintered glass funnel, indicating that no insoluble degradation by-products were formed during the reaction.
- the solution was cooled to 18 °C, and then treated with ethanol (3600 mL) over 80 minutes while agitating the solution at a speed of 400-450 rpm and maintaining the temperature at 20°C. The resulting red slurry was stirred for 30 minutes and then the product collected on a sintered glass funnel.
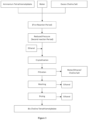
- the reactor was placed under reduced pressure and the evolution of NH 3 monitored by measurement of pH in the gas phase with moist pH indicator paper. When no further evolution of NH 3 was observed (3 hours), 5 L of water was added to the reaction mixture to compensate for loss of water by evaporation and the reaction mixture was transferred to a stainless steel reactor via a cartridge filter. No residues were found on the filter, indicating that no insoluble degradation by-products were formed during the reaction.
- the filtered solution was cooled to 20 °C and then slowly treated with ethanol (80 Kg) over 25 minutes causing crystallisation of the BC-TTM product. Once the addition of ethanol was complete, the resulting slurry was stirred for 30 minutes.
- the final product was then collected on a Hastelloy filter, washed twice with ethanol (2 x 4 Kg) and finally dried at 25°C under vacuum and sieved with a 1.4 mm sieve to afford 3.7 kg of a red crystalline material, 78% yield.
- the BC-TTM product had a purity of 99.2% as assessed by HPLC.
- the reactor was placed under reduced pressure and the evolution of NH 3 monitored by measurement of pH in the gas phase with moist pH indicator paper. When no further evolution of NH 3 was observed (after around 5.5 hours), 16 L of water was added to the reaction mixture to compensate for loss of water by evaporation and the reaction mixture was transferred to a stainless steel reactor via a cartridge filter. No residues were found on the filter, indicating that no insoluble degradation by-products were formed during the reaction.
- the filtered solution was cooled to 20 °C and then slowly treated with ethanol (240 Kg) over 35 minutes causing crystallisation of the product. Once the addition of ethanol was complete, the resulting slurry was stirred for 60 minutes.
- the final product was then collected on a Hastelloy filter, washed twice with ethanol (2 x 10 Kg) and finally dried at 25°C under vacuum and sieved with a 1.4 mm sieve to afford 11.93 kg of a red crystalline material, 80% yield.
- the product had a purity of 99.5% as assessed by HPLC.
- Ammonium tetrathiomolybdate (9 kg) and water (54L) were mixed under stirring to form a dark slurry in a corrosion-resistant jacketed and temperature controlled reactor (Hastelloy), previously purged with N 2 gas and maintained under low N 2 stream. The mixture was cooled to 15 ⁇ 5 °C with cold water.
- the BC-TTM product is isolated on an oyster filter and is then washed twice with ethanol before being recharged to the second reactor for recrystallization.
- the compound in the second reactor is then dissolved in water and transferred to a glass-lined reactor via cartridge filters (0.45 ⁇ m and 0.2 ⁇ m).
- the solution in the glass-lined reactor was heated to 45 ⁇ 3 °C, and then ethanol (undenatured, 95.5%, 304L) was charged over 30 ⁇ 10 minutes.
- the slurry was cooled slowly during at least 60 minutes to 15 ⁇ °C.
- the crystalline product was isolated on a cleaned oyster filter purged with N 2 .
- the solid was washed twice with ethanol, and the oyster filter solid cake was blown dry for 30-60 minutes with N 2 . Drying was performed under reduced pressure at 25 ⁇ 3 °C for at least 8 hours and/or until achieving constant weight.
- the product was then (optionally) sieved and packed. The product is stored in the freezer.
- the resulting crystalline material is a red-orange to dark red or dark burgundy powder with a homogenous appearance.
- X-Ray Powder Diffraction patterns were collected on a Bruker D8 diffractometer (Bruker AXS D8 Advance) using Cu K ⁇ radiation (40 kV, 40 mA), ⁇ - 2 ⁇ goniometer, and divergence of V4 and receiving slits, a Ge monochromator and a Lynxeye detector.
- the instrument is performance checked using a certified Corundum standard (NIST 1976).
- the software used for data collection was Diffrac Plus XRD Commander v2.6.1 and the data were analysed and presented using Diffrac Plus EVA v13.0.0.2 or v15.0.0.0.
- Bis-choline tetrathiomolybdate Form 2 is thermally stable up to about 200°C before degrading.
- Bis-choline tetrathiomolybdate Form 1 was heated from room temperature to 60°C at a rate of 10°C per minute and the crystalline material was isolated.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Acyclic And Carbocyclic Compounds In Medicinal Compositions (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
Claims (15)
- Verfahren zur Herstellung der Verbindung (I):
 Zusammengegeben und Umsetzen von Verbindung (II):
 mit einer ersten Mischung von Verbindung (III):
(NH4)2MoS4 (III)
wobei X-1 für HO-1, Acetat oder ein Halogenid steht; in Wasser, wobei ein molarer Überschuss von Verbindung (II) gegenüber Verbindung (III) verwendet wird,über einen ersten Reaktionszeitraum zur Bereitstellung der ersten Reaktionsmischung;Setzen der ersten Reaktionsmischung unter verminderten Druck über einen zweiten Reaktionszeitraum zur Bereitstellung einer zweiten Reaktionsmischung; undIsolieren von Verbindung (I) aus der zweiten Reaktionsmischung, wobei das Isolieren von Verbindung (I) das Zugeben von Ethanol zu der zweiten Reaktionsmischung umfasst. - Verfahren nach Anspruch 1, wobei X-1 für OH-1 steht.
- Verfahren nach Anspruch 1 oder 2, ferner umfassend das Überwachen der Entwicklung von Ammoniak während des Schritts des Setzens der ersten Reaktionsmischung unter verminderten Druck, beispielsweise durch Messen des pH-Werts von Dampf einer Gasphase in Kontakt mit der ersten Reaktionsmischung, vorzugsweise wobei das Erreichen eines pH-Werts von etwa 7 anzeigt, dass der zweite Reaktionszeitraum abgeschlossen ist.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei das Isolieren von Verbindung (I) das Zugeben von 2 bis 4 Volumina Ethanol zu der zweiten Reaktionsmischung umfasst.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei das Molverhältnis von Verbindung (II) zu Verbindung (III) im Bereich von 4 bis 20, 5 bis 20 oder 5 bis 15 liegt, wobei die gesamte Reaktionszeit des Verfahrens im Vergleich zu einem Verfahren, bei dem Verbindung (II) nicht in einem molaren Überschuss verwendet wird, vorzugsweise um mindestens 50 % verkürzt wird.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei Verbindung (III) sich während des Verfahrens nicht erheblich zersetzt.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei es sich bei der ersten Mischung von Verbindung (III) in Wasser um eine Aufschlämmung von Verbindung (III) in Wasser handelt und/oder eine wässrige Lösung von Verbindung (II) mit der ersten Mischung von Verbindung (III) zusammengegeben wird.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei der Schritt des Zusammengebens von Verbindung (II) mit einer ersten Mischung von Verbindung (III) in Wasser bei einer Temperatur von 10 °C oder weniger, vorzugsweise bei einer Temperatur von 0 °C bis 10 °C, durchgeführt wird.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei der Schritt des Setzens der ersten Reaktionsmischung unter verminderten Druck bei einer Temperatur von 35 °C oder weniger, vorzugsweise bei einer Temperatur von 5 °C bis 35 °C, durchgeführt wird und/oder wobei der Schritt des Setzens der ersten Reaktionsmischung unter verminderten Druck das Setzen der ersten Reaktionsmischung unter einen Druck von 100 mbar oder weniger, vorzugsweise unter einen Druck von 0,1 mbar bis 100 mbar, umfasst.
- Verfahren nach einem der vorhergehenden Ansprüche, wobei der Schritt des Isolierens von Verbindung (I) aus der zweiten Reaktionsmischung das Ausfällen von Verbindung () aus der zweiten Reaktionsmischung und gegebenenfalls das Filtrieren der zweiten Reaktionsmischung zum Erhalt von Verbindung (I) in Form eines Feststoffs umfasst.
- Verfahren nach einem der vorhergehenden Ansprüche, ferner umfassend einen Umkristallisationsschritt gefolgt von fakultativer Polierfiltration, fakultativem Waschen mit Ethanol, Trocknen des erhaltenen Feststoffs und Messen der erhaltenen Teilchengröße des erhaltenen Feststoffs.
- Verfahren nach Anspruch 11, wobei der erhaltene Feststoff ein 50. Perzentil der Teilchengröße (D50) von mindestens 30 µm, mindestens 40 µm, zwischen 30 µm und 90 µm oder zwischen 30 µm und 70 µm aufweist.
- Verfahren nach Anspruch 11 oder 12, wobei der erhaltene Feststoff ein 90. Perzentil der Teilchengröße (D90) zwischen 60 µm und 550 µm, zwischen 400 µm und 550 µm, zwischen 75 µm 135 µm oder zwischen 75 µm und 100 µm aufweist.
- Verfahren nach Anspruch 11, wobei der erhaltene Feststoff einen D50-Wert von weniger als 35 µm und/oder einen D90-Wert von weniger als 75 µm aufweist.
- Verfahren nach Anspruch 11 oder Anspruch 14, wobei der erhaltene Feststoff ein Verhältnis von D90 zu D50 von weniger als 3,0, gegebenenfalls kleiner oder gleich 2,5 oder gegebenenfalls kleiner als 2,0 aufweist.
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US201862630964P | 2018-02-15 | 2018-02-15 | |
| PCT/EP2019/053871 WO2019158719A1 (en) | 2018-02-15 | 2019-02-15 | Process for preparation of bis-choline tetrathiomolybdate |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| EP3752482A1 EP3752482A1 (de) | 2020-12-23 |
| EP3752482B1 true EP3752482B1 (de) | 2023-08-16 |
Family
ID=65516551
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP19706505.5A Active EP3752482B1 (de) | 2018-02-15 | 2019-02-15 | Verfahren zur herstellung von bis-cholin-tetrathiomolybdat |
Country Status (5)
| Country | Link |
|---|---|
| US (1) | US12012424B2 (de) |
| EP (1) | EP3752482B1 (de) |
| JP (1) | JP7394773B2 (de) |
| ES (1) | ES2962600T3 (de) |
| WO (1) | WO2019158719A1 (de) |
Families Citing this family (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN115181407B (zh) * | 2022-08-15 | 2023-12-19 | 万华化学(宁波)有限公司 | 一种无卤阻燃低烟密度聚碳酸酯材料及其制备方法和应用 |
Family Cites Families (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US7189865B2 (en) | 2002-07-23 | 2007-03-13 | Attenuon, Llc | Thiomolybdate analogues and uses thereof |
| WO2017019892A1 (en) | 2015-07-28 | 2017-02-02 | Insmed Incorporated | Compositions comprising copper chelators and methods of use thereof for treating vasculopathies |
| WO2019154876A1 (en) | 2018-02-06 | 2019-08-15 | Wilson Therapeutics Ab | Crystalline particles of bis-choline tetrathiomolybdate |
-
2019
- 2019-02-15 EP EP19706505.5A patent/EP3752482B1/de active Active
- 2019-02-15 JP JP2020543484A patent/JP7394773B2/ja active Active
- 2019-02-15 WO PCT/EP2019/053871 patent/WO2019158719A1/en not_active Ceased
- 2019-02-15 US US16/970,379 patent/US12012424B2/en active Active
- 2019-02-15 ES ES19706505T patent/ES2962600T3/es active Active
Also Published As
| Publication number | Publication date |
|---|---|
| ES2962600T3 (es) | 2024-03-20 |
| JP7394773B2 (ja) | 2023-12-08 |
| US12012424B2 (en) | 2024-06-18 |
| WO2019158719A1 (en) | 2019-08-22 |
| US20200407385A1 (en) | 2020-12-31 |
| JP2021513970A (ja) | 2021-06-03 |
| EP3752482A1 (de) | 2020-12-23 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| KR20210015924A (ko) | 6-(시클로프로판카르복스아미도)-4-((2-메톡시-3-(1-메틸-1h-1,2,4-트리아졸-3-일)페닐)아미노)-n-(메틸-d3) 피리다진-3-카르복스아미드의 결정질 형태 | |
| EP3218351B1 (de) | Verfahren zur herstellung, isolierung und reinigung von pharmazeutisch anwendbaren formen von ahu-377 | |
| EP3141537B1 (de) | Fluorinmarkierungsvorläufer und verfahren zur herstellung einer mit radioaktivem fluorin markierten verbindung mit dieser vorläufer | |
| EP2896609B1 (de) | Kristallines Fingolimod-citrat zur Behandlung von schubförmig auftretender Multipler Sklerose | |
| JP2023062072A (ja) | Rorガンマの阻害剤 | |
| WO2009064479A1 (en) | Polymorphic forms of aliskiren hemifumarate and process for preparation thereof | |
| KR102266680B1 (ko) | 벨리노스테트의 다형태 및 이의 제조 방법 | |
| JP2022163113A (ja) | プリン誘導体を調製するための方法 | |
| EP4077287B1 (de) | Effizientes verfahren zur herstellung von 6-carboxybenzoxazolderivaten | |
| WO2012004391A1 (en) | Process for the preparation of flupirtine maleate | |
| JP7025411B2 (ja) | インドールカルボキサミド化合物の製造方法 | |
| US12012424B2 (en) | Process for preparation of bis-choline tetrathiomolybdate | |
| US20220281836A1 (en) | Process for the preparation of apalutamide | |
| EP3881848B1 (de) | Verfahren zur herstellung von orotsäurederivat | |
| US8278484B2 (en) | Process for preparing a benzoylbenzeneacetamide derivative | |
| WO2010122682A1 (ja) | N-アルコキシカルボニル-tert-ロイシンの製造法 | |
| WO2008096373A2 (en) | Process for synthesizing highly pure nateglinide polymorphs | |
| JP2025528904A (ja) | Tyk2阻害剤およびその使用 | |
| EP3604284B1 (de) | Kristalline form d des eltrombopag-monoethanolamin-salzes | |
| JP4551514B2 (ja) | 不飽和炭化水素残基置換アニリン類の製造方法 | |
| JP2019131532A (ja) | 酸無水物、および該酸無水物を用いたl−カルノシンの製造方法 | |
| JP4564375B2 (ja) | テアニンの製造方法 | |
| KR101925650B1 (ko) | 결정화에 의한 전구체 화합물의 정제 | |
| US20050143431A1 (en) | Novel crystalline forms of parecoxib sodium | |
| JP2026077755A (ja) | クラゾセンタン二ナトリウム塩、その製造及びそれを含有する医薬組成物 |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: UNKNOWN |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: THE INTERNATIONAL PUBLICATION HAS BEEN MADE |
|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: REQUEST FOR EXAMINATION WAS MADE |
|
| 17P | Request for examination filed |
Effective date: 20200914 |
|
| AK | Designated contracting states |
Kind code of ref document: A1 Designated state(s): AL AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HR HU IE IS IT LI LT LU LV MC MK MT NL NO PL PT RO RS SE SI SK SM TR |
|
| AX | Request for extension of the european patent |
Extension state: BA ME |
|
| DAV | Request for validation of the european patent (deleted) | ||
| DAX | Request for extension of the european patent (deleted) | ||
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: EXAMINATION IS IN PROGRESS |
|
| 17Q | First examination report despatched |
Effective date: 20211105 |
|
| GRAP | Despatch of communication of intention to grant a patent |
Free format text: ORIGINAL CODE: EPIDOSNIGR1 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: GRANT OF PATENT IS INTENDED |
|
| INTG | Intention to grant announced |
Effective date: 20230315 |
|
| GRAS | Grant fee paid |
Free format text: ORIGINAL CODE: EPIDOSNIGR3 |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: THE PATENT HAS BEEN GRANTED |
|
| AK | Designated contracting states |
Kind code of ref document: B1 Designated state(s): AL AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HR HU IE IS IT LI LT LU LV MC MK MT NL NO PL PT RO RS SE SI SK SM TR |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: EP |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R096 Ref document number: 602019035051 Country of ref document: DE |
|
| P01 | Opt-out of the competence of the unified patent court (upc) registered |
Effective date: 20230808 |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: DE Ref document number: 602019035051 Country of ref document: DE Ref country code: DE Ref legal event code: R082 Ref document number: 602019035051 Country of ref document: DE Representative=s name: D YOUNG & CO LLP, DE |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R082 Ref document number: 602019035051 Country of ref document: DE Representative=s name: D YOUNG & CO LLP, DE |
|
| REG | Reference to a national code |
Ref country code: LT Ref legal event code: MG9D |
|
| REG | Reference to a national code |
Ref country code: NL Ref legal event code: MP Effective date: 20230816 |
|
| RAP2 | Party data changed (patent owner data changed or rights of a patent transferred) |
Owner name: ALEXION PHARMACEUTICALS, INC. |
|
| REG | Reference to a national code |
Ref country code: AT Ref legal event code: MK05 Ref document number: 1599939 Country of ref document: AT Kind code of ref document: T Effective date: 20230816 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231117 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231216 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: RS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: PT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231218 Ref country code: NO Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231116 Ref country code: NL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: LV Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: LT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: IS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231216 Ref country code: HR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: GR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20231117 Ref country code: FI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: AT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: PL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| REG | Reference to a national code |
Ref country code: ES Ref legal event code: FG2A Ref document number: 2962600 Country of ref document: ES Kind code of ref document: T3 Effective date: 20240320 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SM Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: RO Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: EE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: DK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: CZ Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 Ref country code: SK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R097 Ref document number: 602019035051 Country of ref document: DE |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| 26N | No opposition filed |
Effective date: 20240517 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MC Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LU Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240215 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: CH Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240229 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LU Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240215 Ref country code: CH Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240229 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BG Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R081 Ref document number: 602019035051 Country of ref document: DE Owner name: ALEXION PHARMACEUTICALS, INC., BOSTON, US Free format text: FORMER OWNER: ALEXION PHARMA INTERNATIONAL OPERATIONS UNLIMITED COMPANY, DUBLIN, IE |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BG Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20230816 |
|
| REG | Reference to a national code |
Ref country code: BE Ref legal event code: MM Effective date: 20240229 |
|
| REG | Reference to a national code |
Ref country code: GB Ref legal event code: 732E Free format text: REGISTERED BETWEEN 20241128 AND 20241204 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240229 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240229 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IT Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240215 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: CY Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT; INVALID AB INITIO Effective date: 20190215 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: HU Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT; INVALID AB INITIO Effective date: 20190215 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: GB Payment date: 20260121 Year of fee payment: 8 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: ES Payment date: 20260302 Year of fee payment: 8 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 20260121 Year of fee payment: 8 Ref country code: IE Payment date: 20260123 Year of fee payment: 8 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: FR Payment date: 20260121 Year of fee payment: 8 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: TR Payment date: 20260123 Year of fee payment: 8 |