EP0100877B1 - Procédé de préparation des acides arylacétiques et arylpropioniques - Google Patents
Procédé de préparation des acides arylacétiques et arylpropioniques Download PDFInfo
- Publication number
- EP0100877B1 EP0100877B1 EP83106657A EP83106657A EP0100877B1 EP 0100877 B1 EP0100877 B1 EP 0100877B1 EP 83106657 A EP83106657 A EP 83106657A EP 83106657 A EP83106657 A EP 83106657A EP 0100877 B1 EP0100877 B1 EP 0100877B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- process according
- coordinate
- nicl
- anode
- nickel
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Expired
Links
Images
Classifications
-
- C—CHEMISTRY; METALLURGY
- C25—ELECTROLYTIC OR ELECTROPHORETIC PROCESSES; APPARATUS THEREFOR
- C25B—ELECTROLYTIC OR ELECTROPHORETIC PROCESSES FOR THE PRODUCTION OF COMPOUNDS OR NON-METALS; APPARATUS THEREFOR
- C25B3/00—Electrolytic production of organic compounds
- C25B3/20—Processes
- C25B3/25—Reduction
-
- C—CHEMISTRY; METALLURGY
- C25—ELECTROLYTIC OR ELECTROPHORETIC PROCESSES; APPARATUS THEREFOR
- C25B—ELECTROLYTIC OR ELECTROPHORETIC PROCESSES FOR THE PRODUCTION OF COMPOUNDS OR NON-METALS; APPARATUS THEREFOR
- C25B3/00—Electrolytic production of organic compounds
- C25B3/01—Products
- C25B3/07—Oxygen containing compounds
Definitions
- the invention relates to the preparation of arylacetic and arylpropionic acids from benzyl halides, of formula ArCH Z X and ArCH (CH 3 ) X where Ar denotes a substituted or unsubstituted aromatic group and X a halogen.
- arylacetic and arylpropionic acids are of great importance, since they constitute an essential class of anti-inflammatory drugs, anesthetics and are also precursors in the preparation of penicillins.
- triphenylphosphine P (C 6 H 5 ) 3 was used to form the organic nickel complexes.
- the present invention makes it possible to remedy this drawback and to easily obtain by electro-synthesis of arylacetic and arylpropionic acids.
- It relates to a process for the preparation of arylacetic and arylpropionic acids, comprising an electrochemical reduction, under a carbon dioxide atmosphere, of benzyl halides of formula ArCH 2 X or ArCH (CH 3 ) X, characterized in that said reduction is carried out in the presence of a catalyst comprising at least one organometallic complex derived from a transition metal associated with a bidentate or tetradentate ligand.
- bidentate ligand is meant a ligand which has two coordination sites on the metal used.
- tetradentate ligand is meant a ligand which has four coordination sites on the metal used.
- the transition metal is chosen so that it forms, with the preceding ligands, an electroreducible organometallic complex capable in its reduced form of reacting with the benzyl halide.
- the metal is advantageously chosen from the group comprising nickel and cobalt.
- the organometallic complex is chosen from the group formed on the one hand by nickel bis cyclooctadiene and on the other hand by metallic ligated halides, of formula NiY z L.
- Y being a halogen, L bipyridyl or a diphosphine-type ligand of formula PR 2 - (CH 2 ) n - PR 2 , in which P denotes phosphorus which is a coordination site, R being a radical chosen from the group formed by the phenyl radical and the aliphatic radicals, n being an integer less than or equal to 4.
- n can be equal to 2, 3 or 4.
- diphenyl phosphinoethane (DPPE) of formula:
- diphenyl phosphino propane of formula P (C 6 H 5 ) 2 - (CH 2 ) 3 -P (C 6 H 5 ) 3 or dimethyl phosphino ethane (DMPE) of formula P (CH 3 ) 2 - (CH 2 ) 2 -P (CH 3 ) 2 .
- DPPP diphenyl phosphino propane
- DMPE dimethyl phosphino ethane
- the catalyst consists of a complex M 'salen where M' represents nickel or cobalt and "salen” represents the tetradentate bis salicylidene ethylene diamine ligand, the catalyst having the formula:
- cobalt will be used, which forms with the "saien” ligand a complex more easily electroreducible than the corresponding nickel complex.
- organometallic catalysts according to the invention can be used alone or as a mixture.
- a cocatalyst constituted by a metal ligated halide of formula M 1 Y 2 L ' 2 , L' being a ligand of formula PR ' 3 , R' being chosen from the group formed by the alkyl and aryl radicals, M l being a transition metal, advantageously nickel.
- the second ligand L ' can be used as triphenyl phosphine (TPP) of formula P (C 6 H 5 ) 3 , tributyl phosphine P (C 4 H 9 ) 3 , tricyclohexyl phosphine P (C 6 H 11 ) 3 .
- TPP triphenyl phosphine
- the catalyst used comprises approximately four molar equivalents corresponding to the first complex MY 2 L for a molar equivalent corresponding to the second complex M 1 Y 2 L ' 2 , M being a transition metal.
- the catalyst comprises at least one organometallic complex of the aforementioned type to which a monodentate or bidentate ligand of the aforementioned type is added, for example cyclooctadiene (COD) or bipyridyl.
- COD cyclooctadiene
- bipyridyl for example cyclooctadiene (COD) or bipyridyl.
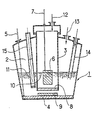
- the single figure very schematically represents an electrolysis cell usable for the implementation of the invention.
- the cell is designated by the reference 1. It comprises two separate compartments, a cathode compartment 2 and an anode compartment 3.
- the cathode 4 can be constituted by a felt, a fabric or a braid of carbon fibers or by a sheet of mercury , with an area of approximately 20 cm 2 .
- the cathode conductor constituted by a copper wire is designated by the reference 5.
- the anode 6 can be of the alterable metal type, lithium, copper, etc., or of the unalterable type, carbon or metal, associated with an oxidizable electrolyte (oxalate for example).
- the anode conductor, consisting of a copper wire, is designated by the reference 7.
- the conductors 5 and 7 are connected to an appropriate generator.
- the reference 8 designates a sintered glass separating the 2 compartments.
- the reference 9 designates a magnetic bar used for agitating the medium.
- the solvent of the electrolyte is formed by a mixture comprising by volume 2/3 of an aprotic solvent, such as tetrahydrofuran (THF), 1/3 of dipolar aprotic solvent such as hexamethylphosphorotriamide (HMPT) or N-methyl pyrrolidone, or tetramethylurea.
- an aprotic solvent such as tetrahydrofuran (THF)
- dipolar aprotic solvent such as hexamethylphosphorotriamide (HMPT) or N-methyl pyrrolidone, or tetramethylurea.
- the electrolyte can be chosen identical or different in the anode 3 and cathode 2 compartments; it is used at a concentration of about 0.1 to 0.3 moles per liter.
- the electrolyte 10 can be of the oxidizable type, advantageously a sodium or lithium oxalate, or non-oxidizable, associated with a soluble anode, for example lithium perchlorate (LiCl0 4 ), tetrabutyl tetrafluoborate ammonium "C4H9) 4NBF4).
- a non-reducible electrolyte 11 (LiCl0 4 , tetrabutylammonium tetrafluoborate) is used in which the benzyl halide and the catalyst according to the invention are introduced.
- a reference electrode 15 consisting of a silver wire immersed in an aprotic solvent solution containing silver perchlorate at a concentration of 0.1 mol / liter, makes it possible to identify the potential of the cathode.
- the arrows 12 and 13 symbolize the introduction, if necessary, of an inert gas into the anode 3 and cathode 2 compartments. Furthermore, carbon dioxide can be introduced into the cathode electrolytic solution by the tube 14 at the atmospheric pressure or slightly higher.
- This elimination can be done for example by adding a magnesium, such as C 2 H S MgX ', X' being a halogen, for example Br, in solution in ether or tetrahydrofuran.
- a magnesium such as C 2 H S MgX ', X' being a halogen, for example Br, in solution in ether or tetrahydrofuran.
- Carbon dioxide is then bubbled through the cathode compartment of the cell, at atmospheric pressure or slightly higher.
- the reaction medium is maintained at ambient temperature or cooled by an external circulation of cold water.
- the electrochemical reduction is then carried out at controlled potential.
- the potential of the agitated mercury sheet compared to the Ag / AgCI0 4 system , is maintained at approximately -2.6 V.
- the electrochemical reduction is carried out until the amount of current passed corresponds to a previously determined value, or until the current is zero.
- the current density at the start of the reduction is approximately 35 mA / cm 2 .
- the solution is then hydrolyzed in an acid medium and extracted with ether.
- the ethereal phase is stirred with aqueous sodium hydroxide and then separated.
- the basic aqueous phase is acidified, saturated with NaCl, then extracted with ether.
- the ethereal phase is dried over MgS0 4 , then evaporated.
- the phenylacetic acid formed is thus recovered, which is characterized by its IR and 1 H NMR spectra and its melting point.
- This step is not necessary if a zerovalent nickel complex such as Ni (COD) 2 is used , but such complexes, which are very oxidizable in air, are less convenient to handle.
- a zerovalent nickel complex such as Ni (COD) 2 is used , but such complexes, which are very oxidizable in air, are less convenient to handle.
- Ni ° L complex is generally very reactive and not very stable. Its stability is increased by the presence of another bidentate ligand in the medium chosen so as to weakly complex Ni ° L, for example COD or bipyridyle, which are relatively weak ligands of zerovalent nickel and do little to hinder its subsequent reaction with the chloride. benzyl.
- This complex can be reduced electrochemically according to:
- This intermediate can evolve by giving dibenzyl C 6 H 5 -CH 2 -CH 2 -C 6 H 5 , but in the presence of C0 2, phenylacetic acid is obtained with regeneration of the zerovalent nickel complex:
- the percentage T1 of C 6 H 5 CH 2 Cl consumed relative to the initial quantity was measured, the percentage RC (chemical yield) of C 6 H 5 CH 2 C00H formed relative to the quantity of C 6 H 5 CH 2 CI consumed, the percentage T3 of C 6 H 5 -CH 2 -CH 2 -C 6 H 5 formed relative to the quantity of initial C 6 H 5 CH 2 Cl, the faradic yield RF, representing the quantity of acid formed relative to the amount of electricity consumed assuming the stoichiometric equation.
- the reaction medium contained 0.1 gram atom of nickel per 1 mole of C 6 H 5 CH 2 Cl, the pressure of C0 2 was 1 atmosphere, the potential was maintained at -2.6 V, unless otherwise stated;
- the solvent of the electrolyte consisted of THF / HMPT (ratio 2/3, 1/3) for examples 1 to 12, (ratio 1/2, 1/2) for examples 13 and 14; for Examples 1 to 9 the electrolyte was 0.1 M LiCl0 4 ; for Examples 10 to 12, the cathode electrolyte was 0.3 M tetrabutylammonium tetrafluoborate, the anode electrolyte being 0.1 M lithium oxalate, with a carbon anode; for examples 13 and 14 the electrolyte was LiCI0 4 0.2 M.
- Example 13 Same conditions as for Example 13 but under two atmospheres of CO 2 .
- the first stage corresponding to the formation of the intermediate complex C 6 H 5 CH 2 NiCIL, operation carried out under neutral gas
- the second to the reduction of this complex in the presence of CO 2 the biaryl derivative is preferably formed.
- the reaction medium contained 0.1 gram atom of nickel per 1 mole of C 6 H 5 CH (CH 3 ) Cl, the pressure of C0 2 was 1 atmosphere, the temperature of 0 °, the potential was maintained at approximately - 2.4, - 2.6 V relative to the Ag + / Ag reference electrode.
- the catalytic species was constituted by Co salen.
- the cathode electrolyte is tetrabutylammonium tetrafluoborate 0.3 M, for example 21, Li ClO 4 0.2 M.
- the solvent of the electrolyte was THF-HMPT (ratio 2/3, 1/3).
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Engineering & Computer Science (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Electrochemistry (AREA)
- Materials Engineering (AREA)
- Metallurgy (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Catalysts (AREA)
- Electrolytic Production Of Non-Metals, Compounds, Apparatuses Therefor (AREA)
Description
- L'invention concerne la préparation des acides arylacétiques et arylpropioniques à partir des halogénures de type benzylique, de formule ArCHZX et ArCH(CH3)X ou Ar désigne un groupe aromatique substitué ou non et X un halogène.
- La fabrication des acides arylacétiques et arylpropioniques revêt une grande importance, car ils constituent une classe essentielle d'anti-inflammatoires, d'anesthésiques et sont également des précurseurs dans la préparation de pénicillines.
- Il est connu de les fabriquer à partir des halogénures de type benzylique par cyanuration, carbonatation ou carbonylation. Toutefois ces réactions sont le plus souvent délicates, de faible sélectivité et de mauvais rendement.
- On sait par ailleurs qu'il est possible de procéder à l'électrosynthèse des acides carboxyliques aromatiques ArC02H à partir d'halogénures aromatiques et de C02 sous pression atmosphérique en utilisant des catalyseurs formés par des complexes organiques de nickel.
- Un tel procédé a été par exemple décrit dans l'article paru dans le Nouveau Journal de Chimie, vol. 5. n° 12 - 1981 pages 621 et suivantes, relatif aux travaux menés par Messieurs TROUPEL, PERICHON et FAUVARQUE et Madame ROLLIN.
- Plus précisément, on a utilisé dans ce procédé, pour former les complexes organiques de nickel, la triphényle phosphine P(C6H5)3.
- Toutefois, on s'est aperçu que le procédé décrit ci-dessus n'était pas directement applicable au cas des halogénures benzyliques car dans ce cas n'était observée que la formation d'un composé bibenzylique. Ainsi dans le cas où le procédé est appliqué au chlorure de benzyle C6H5CH2Cl, on obtient seulement du dibenzyle C6H5-CH2-CH2-C6H5.
- La présente invention permet de remédier à cet inconvénient et d'obtenir facilement par électro- synthèse des acides arylacétiques et arylpropioniques.
- Elle a pour objet un procédé de préparation des acides arylacétiques et arylpropioniques, comportant une réduction électrochimique, sous atmosphère de dioxyde de carbone, d'halogénures de type benzylique de formule ArCH2X ou ArCH(CH3)X, caractérisé par le fait que ladite réduction est effectuée en présence d'un catalyseur comprenant au moins un complexe organométallique dérivé d'un métal de transition associé à un coordinat bidenté ou tétradenté.
- Par coordinat bidenté on désigne un ligand qui a deux sites de coordination sur le métal utilisé. Par coordinat tétradenté, on désigne un ligand qui a quatre sites de coordination sur le métal utilisé.
- Le métal de transition est choisi de telle sorte qu'il forme avec les coordinats précédents un complexe organométallique électroréductible capable sous sa forme réduite de réagir avec l'halogénure de type benzylique. Le métal est avantageusement choisi dans le groupe comprenant le nickel et le cobalt.
- Selon l'invention, le complexe organométallique est choisi dans le groupe formé d'une part par le nickel bis cyclooctadiène et d'autre part par les halogénures ligandés métalliques, de formule NiYzL. Y étant un halogène, L le bipyridyle ou un coordinat de type diphosphine de formule PR2-(CH2)n- PR2, dans laquelle P désigne le phosphore qui est un site de coordination, R étant un radical choisi dans le groupe formé par le radical phényle et les radicaux aliphatiques, n étant un nombre entier inférieur ou égal à 4.
- Lorsque R est le radical phényle, n peut être égal à 2, 3 ou 4. Lorsque R désigne le radical méthyle, avantageusement n = 2.
- Selon un mode de réalisation de l'invention, on utilise comme coordinat L le diphényle phosphino éthane (DPPE) de formule :

- On peut également utiliser le diphényle phosphino propane (DPPP) de formule P(C6H5)2-(CH2)3-P(C6H5)3 ou le diméthyle phosphino éthane (DMPE) de formule P(CH3)2-(CH2)2-P(CH3)2.
- Selon un autre mode de réalisation de l'invention le catalyseur est constitué par un complexe M' salen où M' représente le nickel ou le cobalt et « salen » représente le coordinat tétradenté bis salicylidène éthylène diamine, le catalyseur ayant la formule :

- Avantageusement, on utilisera le cobalt qui forme avec le coordinat « saien un complexe plus facilement électroréductible que le complexe correspondant du nickel.
- Les catalyseurs organométalliques conformes à l'invention peuvent être utilisés seuls ou en mélange.
- On peut également leur ajouter un cocatalyseur constitué par un halogénure ligandé métallique de formule M1Y2L'2, L' étant un coordinat de formule PR'3, R' étant choisi dans le groupe formé par les radicaux alkyle et aryle, Ml étant un métal de transition, avantageusement le nickel.
- Ainsi, on peut utiliser comme second coordinat L' la triphényle phosphine (TPP) de formule P(C6H5)3, la tributyle phosphine P(C4H9)3, la tricyclohexyle phosphine P(C6H11)3.
- Avantageusement, le catalyseur utilisé comporte environ quatre équivalents molaires correspondant au premier complexe MY2L pour un équivalent molaire correspondant au second complexe M1Y2L'2, M étant un métal de transition.
- Selon une autre caractéristique de l'invention, le catalyseur comprend au moins un complexe organométallique du type précité auquel on ajoute un coordinat monodenté ou bidenté du type précité, par exemple le cyclooctadiène (COD) ou le bipyridyle.
- D'autres caractéristiques de l'invention ressortiront de la description qui va suivre relative à différents exemples de mise en oeuvre de l'invention.
- La figure unique représente très schématiquement une cellule d'électrolyse utilisable pour la mise en oeuvre de l'invention.
- La cellule est désignée par la référence 1. Elle comprend deux compartiments séparés, un compartiment cathodique 2 et un compartiment anodique 3. La cathode 4 peut être constituée par un feutre, un tissu ou une tresse de fibres de carbone ou par une nappe de mercure, de surface environ 20 cm2. Le conducteur cathodique constitué par un fil de cuivre, est désigné par la référence 5.
- L'anode 6 peut être du type métal altérable, lithium, cuivre, etc., ou du type inaltérable, carbone ou métal, associé à un électrolyte oxydable (oxalate par exemple). Le conducteur anodique, constitué par un fil de cuivre, est désigné par la référence 7.
- En vue de la réduction électrochimique les conducteurs 5 et 7 sont reliés à un générateur approprié.
- La référence 8 désigne un verre fritté séparant les 2 compartiments.
- La référence 9 désigne une barre aimantée servant à l'agitation du milieu.
- Le solvant de l'électrolyte est formé par un mélange comprenant en volume 2/3 d'un solvant aprotique, tel que le tétrahydrofuranne (THF), 1/3 de solvant aprotique dipolaire tel que l'hexaméthylphos- phorotriamide (HMPT) ou la N-méthyl pyrrolidone, ou la tétraméthylurée.
- L'électrolyte peut être choisi identique ou différent dans les compartiments anodique 3 et cathodique 2 ; il est utilisé à la concentration d'environ 0,1 à 0,3 mole par litre. Ainsi dans le compartiment anodique 3 l'électrolyte 10 peut être du type oxydable, avantageusement un oxalate de sodium ou de lithium, ou non oxydable, associé à une anode soluble, par exemple du perchlorate de lithium (LiCI04), du tétrafluoborate de tétrabutyle ammonium «C4H9)4NBF4).
- Dans le compartiment cathodique 2, on utilise un électrolyte 11 non réductible (LiCI04, tétrabutylammonium tétrafluoborate) dans lequel on introduit l'halogénure de type benzylique et le catalyseur conforme à l'invention.
- Une électrode de référence 15, constituée d'un fil d'argent immergé dans une solution de solvant aprotique contenant du perchlorate d'argent à la concentration 0,1 mole/litre, permet de repérer le potentiel de la cathode.
- Les flèches 12 et 13 symbolisent l'introduction, si nécessaire, d'un gaz inerte dans les compartiments anodique 3 et cathodique 2. Par ailleurs, on peut introduire, dans la solution électrolytique cathodique, du gaz carbonique, par le tube 14 à la pression atmosphérique ou légèrement supérieure.
- De manière à éviter des réactions secondaires, on élimine soigneusement l'eau résiduelle contenue dans le milieu électrolytique.
- Cette élimination peut se faire par exemple par addition d'un magnésien, tel que C2HSMgX', X' étant un halogène, par exemple Br, en solution dans l'éther ou le tétrahydrofuranne.
- Pour la préparation de l'acide phénylacétique C6H5CH2CO2H, on introduit 5 milli-moles de chlorure de benzyle C6H5CH2Cl dans le compartiment cathodique 2. On ajoute également le catalyseur conforme à l'invention en quantité telle qu'à une mole de chlorure de benzyle corresponde 0,1 atome-gramme de métal de transition.
- On fait ensuite barboter du dioxyde de carbone dans le compartiment cathodique de la cellule, à la pression atmosphérique ou légèrement supérieure.
- Le milieu réactionnel est maintenu à température ambiante ou refroidi par une circulation extérieure d'eau froide.
- La réduction électrochimique est ensuite effectuée à potentiel contrôlé.
- Ainsi, le potentiel de la nappe de mercure agitée, par rapport au système Ag/AgCI04, est maintenue à environ -2,6 V.
- La réduction électrochimique est effectuée jusqu'à ce que la quantité de courant passée corresponde à une valeur déterminée préalablement, ou jusqu'à ce que le courant soit nul.
- La densité de courant au début de la réduction est d'environ 35 mA/cm2.
- La solution est ensuite hydrolysée en milieu acide et extraite à l'éther.
- La phase éthérée est agitée avec de la soude aqueuse puis séparée.
- L'analyse par chromatographie en phase vapeur de la phase éthérée permet de calculer la quantité de C6H5CH2Cl restante, ainsi que la quantité de C6H5-CH2-CH2-C6H5 formée.
- La phase aqueuse basique est acidifiée, saturée en NaCI, puis extraite à l'éther. La phase éthérée est séchée sur MgS04, puis évaporée.
- On récupère ainsi l'acide phénylacétique formé, qui est caractérisé par ses spectres I.R. et R.M.N.1H et son point de fusion.
- Le principe de la méthode, exposé pour la fabrication de l'acide phénylacétique à partir de chlorure de benzyle, en présence d'un halogénure ligandé de nickel NiY2L est le suivant :
- Dans un premier temps, on forme électrochimiquement un complexe intermédiaire par insertion du métal de transition, par exemple le nickel, au sein de la liaison C-CI du chlorure de benzyle.
- On a dans un premier temps la réaction :

- Cette étape n'est pas nécessaire si l'on utilise un complexe de nickel zérovalent tel que Ni(COD)2, mais de tels complexes, très oxydables à l'air, sont de manipulation moins commode.
- Le complexe Ni°L est en général très réactif et peu stable. Sa stabilité est augmentée par la présence d'un autre coordinat bidenté dans le milieu choisi de façon à complexer faiblement Ni°L, par exemple COD ou bipyridyle, qui sont des coordinats relativement faibles du nickel zérovalent et gênent peu sa réaction ultérieure avec le chlorure de benzyle.
- Dans un deuxième temps on a :

- Le bilan global étant

- Ce complexe peut être réduit électrochimiquement selon :

- Cet intermédiaire peut évoluer en donnant du dibenzyle C6H5-CH2-CH2-C6H5, mais en présence de C02 on obtient l'acide phénylacétique avec régénération du complexe de nickel zérovalent :
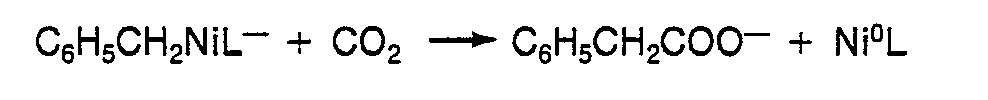
- Le cycle catalytique peut alors se poursuivre. Globalement on a la réaction :

- Les réactions sont analogues à partir d'autres complexes organométalliques conformes à l'invention.
- Plusieurs exemples de préparation ont été effectués à partir de C6H5CH2Cl en modifiant la nature de l'espèce catalytique, ainsi que la température du milieu.
- Pour ces exemples, on a mesuré le pourcentage T1 de C6H5CH2Cl consommé par rapport à la quantité initiale, le pourcentage RC (rendement chimique) de C6H5CH2C00H formé par rapport à la quantité de C6H5CH2CI consommé, le pourcentage T3 de C6H5-CH2-CH2-C6H5 formé par rapport à la quantité de C6H5CH2Cl initiale, le rendement faradique RF, représentant la quantité d'acide formée rapportée à la quantité d'électricité consommée en supposant l'équation stcechiométrique.
- Pour tous ces exemples, le milieu réactionnel comportait 0,1 atome-gramme de nickel pour 1 mole de C6H5CH2Cl, la pression de C02 était de 1 atmosphère, le potentiel était maintenu à - 2,6 V, sauf mention contraire ; le solvant de l'électrolyte était constitué par THF/HMPT (rapport 2/3, 1/3) pour les exemples 1 à 12, (rapport 1/2, 1/2) pour les exemples 13 et 14 ; pour les exemples 1 à 9 l'électrolyte était LiCI04 0,1 M ; pour les exemples 10 à 12, l'électrolyte cathodique était le tétrabutylammonium tétrafluoborate 0,3 M, l'électrolyte anodique étant l'oxalate de lithium 0,1 M, avec une anode en carbone ; pour les exemples 13 et 14 l'électrolyte était LiCI04 0,2 M.
- Espèce catalytique
- NiCl2, DPPE et NiCl2 (TPP)2 dans un rapport molaire de 4/1
- Température 20 °C
- Arrêt de l'électrolyse à courant nul.
- Espèce catalytique
- NiCl2, DPPE et NiCl2 (TPP)2 dans un rapport molaire de 4/1
- Arrêt de l'électrolyse après 8 heures.
- On constate qu'à 0 °C l'électrolyse est beaucoup plus lente qu'à 20 °C.
- Espèce catalytique
- NiCl2, DPPE et NiCl2 (TPP)2 dans un rapport molaire de 19/1
- Arrêt de l'électrolyse après 8 heures.
- Espèce catalytique
- NiCl2, DPPE et NiCl2 (TPP)2 dans un rapport molaire de 19/1
- Arrêt de l'électrolyse après 15 heures.
- Espèce catalytique
- NiCl2, DMPE et NiCl2 (TPP)2 dans un rapport molaire de 4/1
- Arrêt de l'électrolyse lorsque le courant devient trop faible.
- Espèce catalytique
- NiCl2, DMPE et NiCl2 (TPP)2 dans un rapport molaire de 19/1
- Espèce catalytique
- NiCl2, DPPP et NiCl2 (TPP)2 dans un rapport molaire de 4/1
- Espèce catalytique
- NiCl2, DPPE et NiCl2, [P(C6H11)3]2 dans un rapport molaire de 4/1
- Espèce catalytique
- NiCl2, DPPE
- Espèce catalytique
- NiCl2, DPPP + COD en proportion molaire 1/1
- Electrolyse terminée en 5 heures.
- Espèce catalytique
- Nickel bis cyclooctadiène
- Electrolyse arrêtée au bout de 20 heures.
- Espèce catalytique
- NiCl2, bipyridyle
- Electrolyse pendant 25 heures.
- Espèce catalytique
- Cobalt salen
- Electrolyse à -2,3 V sur cathode de mercure au potentiel de réduction de Co salen ; conversion totale en 20 heures.
- Mêmes conditions que pour l'exemple 13 mais sous deux atmosphères de CO2.
- Les résultats des mesures sont explicités sur le tableau suivant :


- On a pu constater qu'il était souhaitable de réaliser la réduction électrochimique en une seule étape.
- En effet, si l'on procède en deux temps, le premier temps correspondant à la formation du complexe intermédiaire C6H5CH2NiCIL, opération réalisée sous gaz neutre, le deuxième à la réduction de ce complexe en présence de CO2, il se forme préférentiellement le dérivé biarylique.
- Ainsi si l'on reprend les conditions du 1er exemple en effectuant une première réduction électrochimique sous argon à un potentiel de - 2,1 V, suivie d'une seconde réduction en présence de C02 à un potentiel de - 2,6 V, la valeur obtenue pour T3 passe de 18 à 48.
- On a également réalisé des exemples correspondant à la préparation d'acides arylpropioniques.
- On a ainsi procédé à la synthèse de l'acide phényle propionique C6H5CH(CH3)COOH à partir de C6H5CH(CH3)Cl.
- En raison de la structure de ce composé, il existe une réaction parasite supplémentaire, conduisant par élimination de HCl à la formation de styrène. Pour éviter cette réaction il est avantageux d'une part de conduire l'opération à une température inférieure à la température ambiante, par exemple à 0 °C, d'autre part d'ajouter lorsque le catalyseur est NiY2L un bidenté supplémentaire, COD ou bipyridyle par exemple, qui se coordonne faiblement au nickel zérovalent.
- L'utilisation de Co salen comme catalyseur ne nécessite pas de coordinat supplémentaire.
- Plusieurs exemples de préparation ont été effectués à partir de C6H5―CH(CH3)Cl.
- Pour tous ces exemples, on a mesuré le pourcentage T'1 de C6H5CH(CH3)Cl consommé par rapport à la quantité initiale, le rendement chimique RC' et le rendement faradique RF'. Le produit secondaire est le styrène. On a la réaction globale :

- Pour les exemples 15 à 20, le milieu réactionnel comportait 0,1 atome-gramme de nickel pour 1 mole de C6H5CH(CH3)Cl, la pression de C02 était de 1 atmosphère, la température de 0°, le potentiel était maintenu à environ - 2,4, - 2,6 V par rapport à l'électrode de référence Ag+/Ag. Pour l'exemple 21, l'espèce catalytique était constituée par Co salen.
- Pour les exemples 15 à 20 et l'électrolyte cathodique est le tétrabutylammonium tétrafluoborate 0,3 M, pour l'exemple 21, Li ClO4 0,2 M. Le solvant de l'électrolyte était THF-HMPT (rapport 2/3, 1/3).
- Espèce catalytique
- NICl2, DPPP
- T1' : 40, RC' : 57, RF' : 73.
- Espèce catalytique
- NiCl2, DPPE + COD DPPE et COD dans un rapport molaire de 1/1,
- T1' : 72 RC' : 82, RF' : 74.
- Espèce catalytique
- NiClz, DPPP + COD DPPP et COD dans un rapport molaire de 1/1,
- RC' : 71 RF' : 94
- Espèce catalytique
- NiCl2, DPPP + bipyridyle, DPPP et bipyridyle dans un rapport molaire de 1/1,
- T1' : 82, RC' : 51, RF' : 44
- Espèce catalytique
- NiClz, DPPP + COD, DPPP et COD dans un rapport molaire de 1/1,
- T1' : 100 RC' : 75 RF' : 75
- Espèce catalytique
- NiCl2, DPPP + COD DPPP et COD dans un rapport molaire de 1/1,
- T1' : 96 RC' : 89 RF' : 93
- Espèce catalytique
- Co salen
- T1' : 100 RC' : 60
- Le procédé décrit ci-dessus peut ainsi être directement appliqué à la synthèse d'un anti inflammatoire commercial, le naproxène, selon la réaction

- Espèce catalytique
- NiC12, DPPP + COD DPPP et COD dans un rapport molaire de 1/1 à 0 °C
- T1':100 RC':66 RF':66
- Bien entendu l'invention n'est nullement limitée aux modes d'exécution qui n'ont été donnés qu'à titre d'exemple.
Température 0 °C
Température 20 °C
Température 0 °C
Température 20 °C
Mêmes conditions que l'exemple 5
Mêmes conditions que l'exemple 5.
Mêmes conditions que l'exemple 5.
Mêmes conditions que l'exemple 5.
Température 20 °C
Température 20 °C
Température 20 °C
CO2 sous pression atmosphérique
anode en cuivre
électrolyse jusqu'à courant nul
anode en cuivre
électrolyse en 20 heures
anode en cuivre,
arrêt de l'électrolyse à 55 % de la quantité d'électricité théorique
anode de cuivre
anode de platine,
électrolyte anodique oxalate de sodium, 0,1 M
cathode fibres de carbone tressées et non plus mercure
anode en platine, électrolyte anodique : oxalate de lithium
électrolyse complète en 12 heures
CO2 sous 1 atmosphère
électrolyse à - 2 volts à 20 °C
Claims (19)
Priority Applications (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| AT83106657T ATE20480T1 (de) | 1982-07-13 | 1983-07-07 | Verfahren zur herstellung von arylessig- und arylpropionsaeure. |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| FR8212275A FR2530266B1 (fr) | 1982-07-13 | 1982-07-13 | Procede de preparation des acides arylacetiques et arylpropioniques |
| FR8212275 | 1982-07-13 |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| EP0100877A1 EP0100877A1 (fr) | 1984-02-22 |
| EP0100877B1 true EP0100877B1 (fr) | 1986-06-18 |
Family
ID=9275950
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP83106657A Expired EP0100877B1 (fr) | 1982-07-13 | 1983-07-07 | Procédé de préparation des acides arylacétiques et arylpropioniques |
Country Status (6)
| Country | Link |
|---|---|
| US (1) | US4517061A (fr) |
| EP (1) | EP0100877B1 (fr) |
| JP (1) | JPS5935688A (fr) |
| AT (1) | ATE20480T1 (fr) |
| DE (1) | DE3364177D1 (fr) |
| FR (1) | FR2530266B1 (fr) |
Families Citing this family (20)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| FR2566434B1 (fr) * | 1984-06-21 | 1986-09-26 | Poudres & Explosifs Ste Nale | Procede d'electrosynthese d'acides carboxyliques |
| FR2579627B1 (fr) * | 1985-03-29 | 1987-05-15 | Poudres & Explosifs Ste Nale | Procede d'electrosynthese d'alcools |
| IT1216929B (it) * | 1987-04-16 | 1990-03-14 | Enichem Sintesi | Procedimento per la sintesi di acidi 2-aril-propionici. |
| JPS63295021A (ja) * | 1987-05-28 | 1988-12-01 | Nippon Steel Corp | 溶接管における薄肉管成形法 |
| DE19858208B4 (de) * | 1998-12-17 | 2004-03-04 | Fhp Motors Gmbh | Schutzleiteranschluß an einem lamellierten Ständerblechpaket eines Elektromotors |
| GB0228018D0 (en) * | 2002-11-30 | 2003-01-08 | Lucite Int Uk Ltd | Carbonylation of ester |
| GB0315536D0 (en) * | 2003-07-03 | 2003-08-06 | Lucite Int Uk Ltd | Process for the hydroformylation of ethylenically unsaturated compounds |
| WO2005003070A1 (fr) * | 2003-07-03 | 2005-01-13 | Lucite International Uk Limited | Procede d'hydroformylation de composes ethyleniquement insatures |
| GB0403592D0 (en) * | 2004-02-18 | 2004-03-24 | Lucite Int Uk Ltd | A catalyst system |
| EP1732531A2 (fr) | 2004-02-25 | 2006-12-20 | La Jolla Pharmaceutical Company | Amines et amides pour le traitement de maladies |
| GB0411951D0 (en) * | 2004-05-28 | 2004-06-30 | Lucite Int Uk Ltd | Carbonylation of ester |
| GB0516556D0 (en) * | 2005-08-12 | 2005-09-21 | Lucite Int Uk Ltd | Improved catalyst system |
| CA2626107C (fr) | 2005-11-17 | 2016-06-21 | Lucite International Uk Limited | Carbonylation de composes a insaturations ethyleniques |
| GB0607494D0 (en) | 2006-04-13 | 2006-05-24 | Lucite Int Uk Ltd | Metal complexes |
| MX2009005568A (es) * | 2006-12-02 | 2009-06-05 | Lucite Int Uk Ltd | Nuevos ligandos de carbonilacion y su uso en la carbonilacion de compuestos etilenicamente insaturados. |
| GB0625518D0 (en) * | 2006-12-21 | 2007-01-31 | Lucite Int Uk Ltd | Carbonylation of conjugated dienes |
| GB0812297D0 (en) * | 2008-07-04 | 2008-08-13 | Lucite Int Uk Ltd | Novel carbonylation ligand sand thier use of in the carbonylation of ethylenically unsaturated compounds |
| GB201000078D0 (en) | 2010-01-05 | 2010-02-17 | Lucite Int Uk Ltd | Process for the carbonylation of ethylenically unsaturated compounds, novel carbonylation ligands and catalyst systems incorporatng such ligands |
| JP5842774B2 (ja) * | 2012-09-20 | 2016-01-13 | コニカミノルタ株式会社 | 光電変換素子および太陽電池 |
| CN113136593A (zh) * | 2021-04-14 | 2021-07-20 | 赤峰学院 | 一种布洛芬的合成方法 |
Family Cites Families (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US3344045A (en) * | 1964-10-23 | 1967-09-26 | Sun Oil Co | Electrolytic preparation of carboxylic acids |
| US3764492A (en) * | 1972-01-10 | 1973-10-09 | Monsanto Co | Electrolytic preparation of esters from organo halides |
-
1982
- 1982-07-13 FR FR8212275A patent/FR2530266B1/fr not_active Expired
-
1983
- 1983-07-07 AT AT83106657T patent/ATE20480T1/de active
- 1983-07-07 EP EP83106657A patent/EP0100877B1/fr not_active Expired
- 1983-07-07 DE DE8383106657T patent/DE3364177D1/de not_active Expired
- 1983-07-12 JP JP58126822A patent/JPS5935688A/ja active Granted
- 1983-07-13 US US06/513,497 patent/US4517061A/en not_active Expired - Fee Related
Also Published As
| Publication number | Publication date |
|---|---|
| DE3364177D1 (en) | 1986-07-24 |
| FR2530266A1 (fr) | 1984-01-20 |
| JPH0229755B2 (fr) | 1990-07-02 |
| EP0100877A1 (fr) | 1984-02-22 |
| ATE20480T1 (de) | 1986-07-15 |
| FR2530266B1 (fr) | 1985-07-12 |
| US4517061A (en) | 1985-05-14 |
| JPS5935688A (ja) | 1984-02-27 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP0100877A1 (fr) | Procédé de préparation des acides arylacétiques et arylpropioniques | |
| Derien et al. | From stoichiometry to catalysis: electroreductive coupling of alkynes and carbon dioxide with nickel-bipyridine complexes. Magnesium ions as the key for catalysis | |
| Derien et al. | Activation of carbon dioxide: nickel-catalyzed electrochemical carboxylation of diynes | |
| JP2755327B2 (ja) | α−飽和ケトンの電気化学的合成方法 | |
| Garnier et al. | Electrosynthesis of symmetrical ketones from organic halides and carbon dioxide catalysed by 2, 2′-bipyridine-nickel complexes | |
| GB2230782A (en) | Process for the electrosynthesis of beta, gamma-unsaturated esters | |
| CA1272180A (fr) | Systeme catalytique, son procede de preparation et son application a la fabrication d'aldehydes | |
| FR2725443A1 (fr) | Preparation d'acides carboxyliques ou des esters correspondants par carbonylation en presence d'iridium | |
| EP0955303A2 (fr) | Complexe ruthénium-iodo-phosphine optiquement actif | |
| Jutand et al. | Activation of Aryl and Vinyl Triflates by Palladium and Electron Transfer–Electrosynthesis of Aromatic and α, β‐Unsaturated Carboxylic Acids from Carbon Dioxide | |
| Oçafrain et al. | Electrochemical generation of a nickel–carbonyl complex, catalyst for the electroreductive coupling of organic halides and carbon monoxide into ketones | |
| Davies et al. | Electrochemical generation and reactivity of bis (tertiary phosphine) platinum (0) complexes: a comparison of the reactivity of [Pt (PPh3) 2] and [Pt (PEt3) 2] equivalents | |
| EP0201365B1 (fr) | Procédé d'électrosynthèse d'alcools et de composés époxydes | |
| Bedioui et al. | Electroassisted catalysis of the reductive coupling of 2-bromooctane and methyl vinyl ketone by a binuclear cobalt–salen–iron complex in DMF solution: electrosynthesis and cyclic voltammetry analysis | |
| Olivero et al. | Selectivity in the tandem cyclization–carboxylation reaction of unsaturated haloaryl ethers catalyzed by electrogenerated nickel complexes | |
| US20030162994A1 (en) | 6,6'-Bis-(1-Phosphanorbornadiene) diphosphines, their preparation and their uses | |
| EP0196253B1 (fr) | Système catalytique, son procédé de préparation et son application à la fabrication d'aldéhydes | |
| WO2013114311A1 (fr) | Procede de preparation de phosphines tertiaires et de derives de celles-ci | |
| CH664979A5 (fr) | Procede d'electrosynthese d'acides carboxyliques. | |
| JP2009215647A (ja) | 炭酸ジエステルの製造方法 | |
| Gloaguen et al. | Electrochemistry of dinuclear, thiolate-bridged transition-metal compounds. 7. Electrochemical generation of isomers of [Mo2Cp2 (CO) 2 (. mu.-SMe) 2] and their reactivity toward isocyanide ligands | |
| Chiarotto et al. | Palladium-catalysed electrochemical reduction of vinyl and aryl triflates | |
| Sarkar et al. | Room Temperature Electroreductive Carboxylation of Unactivated Alkyl Chlorides with CO2 Using a Ni (II) NNN-Pincer Complex. Towards Polyvinyl chloride (PVC) Upgrading | |
| Budnikova et al. | Electrochemical synthesis of phosphorus esters from white phosphorus in the presence of copper complexes and ethanol | |
| EP1245551B1 (fr) | 1,2-bis(methyl(1,1,3,3-tetramethylbutyl)-phosphino)ethane, son procede de preparation, complexes de metaux de transition le contenant en tant que ligand et leurs utilisations |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| AK | Designated contracting states |
Designated state(s): AT BE CH DE FR GB IT LI LU NL SE |
|
| 17P | Request for examination filed |
Effective date: 19840809 |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| AK | Designated contracting states |
Kind code of ref document: B1 Designated state(s): AT BE CH DE FR GB IT LI LU NL SE |
|
| REF | Corresponds to: |
Ref document number: 20480 Country of ref document: AT Date of ref document: 19860715 Kind code of ref document: T |
|
| REF | Corresponds to: |
Ref document number: 3364177 Country of ref document: DE Date of ref document: 19860724 |
|
| ITF | It: translation for a ep patent filed | ||
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| 26N | No opposition filed | ||
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: LU Payment date: 19910607 Year of fee payment: 9 |
|
| EPTA | Lu: last paid annual fee | ||
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: SE Payment date: 19920522 Year of fee payment: 10 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: CH Payment date: 19920526 Year of fee payment: 10 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: FR Payment date: 19920527 Year of fee payment: 10 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 19920615 Year of fee payment: 10 Ref country code: BE Payment date: 19920615 Year of fee payment: 10 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: GB Payment date: 19920625 Year of fee payment: 10 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LU Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 19920707 |
|
| ITTA | It: last paid annual fee | ||
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: NL Payment date: 19920731 Year of fee payment: 10 Ref country code: AT Payment date: 19920731 Year of fee payment: 10 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GB Effective date: 19930707 Ref country code: AT Effective date: 19930707 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SE Effective date: 19930708 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LI Effective date: 19930731 Ref country code: CH Effective date: 19930731 Ref country code: BE Effective date: 19930731 |
|
| BERE | Be: lapsed |
Owner name: CENTRE NATIONAL DE LA RECHERCHE SCIENTIFIQUE CNRS Effective date: 19930731 Owner name: CIE GENERALE D'ELECTRICITE Effective date: 19930731 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: NL Effective date: 19940201 |
|
| GBPC | Gb: european patent ceased through non-payment of renewal fee |
Effective date: 19930707 |
|
| NLV4 | Nl: lapsed or anulled due to non-payment of the annual fee | ||
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: FR Effective date: 19940331 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Effective date: 19940401 |
|
| REG | Reference to a national code |
Ref country code: FR Ref legal event code: ST |
|
| EUG | Se: european patent has lapsed |
Ref document number: 83106657.6 Effective date: 19940210 |