WO2009082913A1 - Process for isolation of a mixture of rrrs and sssr configurations of nebivolol intermediates - Google Patents
Process for isolation of a mixture of rrrs and sssr configurations of nebivolol intermediates Download PDFInfo
- Publication number
- WO2009082913A1 WO2009082913A1 PCT/CN2008/073492 CN2008073492W WO2009082913A1 WO 2009082913 A1 WO2009082913 A1 WO 2009082913A1 CN 2008073492 W CN2008073492 W CN 2008073492W WO 2009082913 A1 WO2009082913 A1 WO 2009082913A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- solvent
- sssr
- rrrs
- mixture
- formula
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D311/00—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings
- C07D311/02—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings ortho- or peri-condensed with carbocyclic rings or ring systems
- C07D311/04—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring
- C07D311/58—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring other than with oxygen or sulphur atoms in position 2 or 4
Definitions
- the present invention relates to a process for the preparation of a mixture of isomers of a compound, and in particular to a process for the preparation of a mixture of nebivolol intermediates of the RRRS and SSSR types.
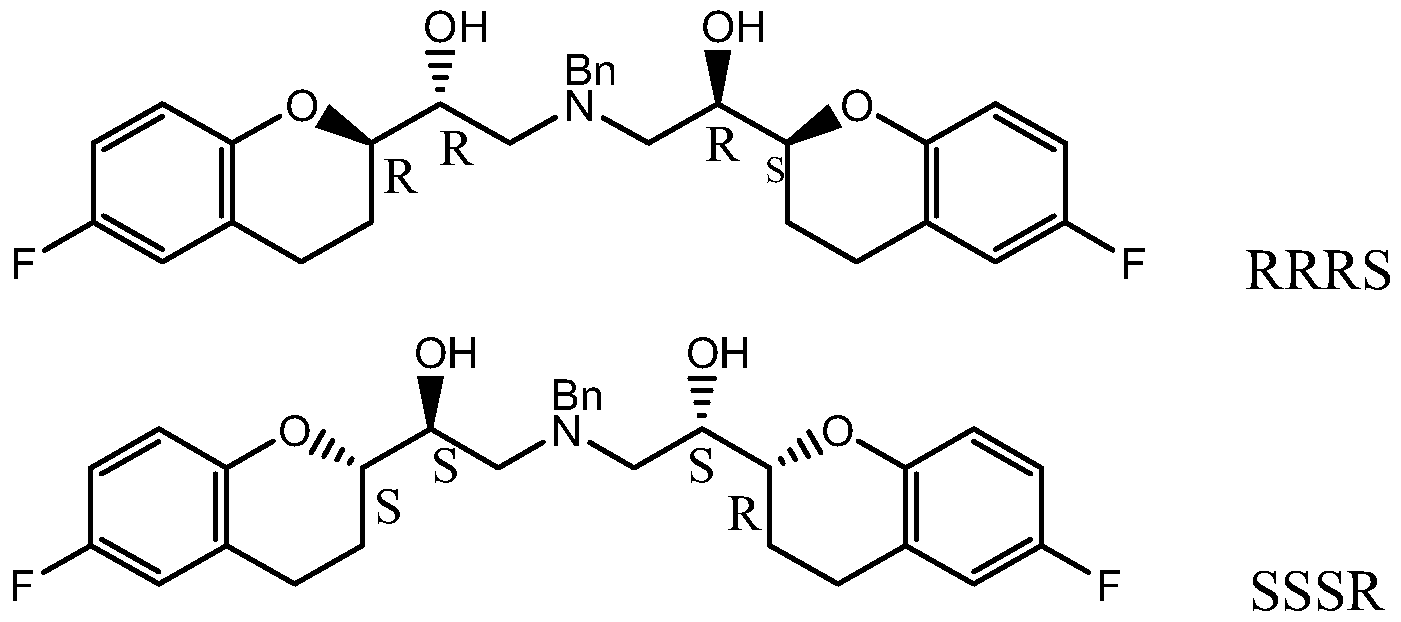
- Nebivolol has the following formula, which has four chiral carbon atoms in its structure, and theoretically has 16 stereoisomers. Considering the symmetry factor, only 10 stereoisomers exist.
- d-naprolol RRRS
- 1-naprozol SSSR
- the synthetic route is as follows;
- 8a is the most critical intermediate for the preparation of nebivolol, not only because 8a is debenzylated to obtain nebivolol, but also by directly controlling the stereo configuration of 8a to obtain the desired naibi Lol stereoisomers.
- the obtained 8a is a mixture of (RRRS, SSSR, RRSR, SSRS):
- WO2006016376 reports that 8a is converted to the hydrochloride salt, which is recrystallized and then freed to give the desired stereoisomer. The process is abruptly long and the yield is low. WO2007083318 improves this. After the formation of 8a, the solvent ethanol is evaporated to dryness, and isopropyl ether is added. The obtained solid is treated with isopropyl ether. After two treatments, 2% of other isomers are still present, and the subsequent operations are carried out. bring inconvenience. Summary of the invention
- the technical problem to be solved by the present invention is to overcome the existing method of synthesizing RRRS and SSSR type nebivolol, and the key steps of separating the intermediate mixture of RRRS, SSSR, RRSR and SSRS type are inconvenient and the separation efficiency is low.
- the defect provides a simple and highly efficient separation method to prepare a mixture of the Nebivolol intermediates of the RRRS and SSSR types.
- the method of the present invention can be any of the following two ways:
- X is hydrogen, or a sulfhydryl group of Cr ⁇ Qs or ( ⁇ ( 6 methoxyl, n is 1 to 5; preferably X is hydrogen).
- the alcohol solvent is preferably one or more of methanol, ethanol, propanol and isopropanol, more preferably ethanol.
- the precipitating solvent is preferably a C 5 - C 9 hydrocarbon solvent or an ether solvent, including a cyclic hydrocarbon, preferably a ruthenium, an n-glyoxime and/or a cyclohexane, more preferably It is already awkward.
- the precipitation solvent may also preferably be an ether solvent, more preferably isopropyl ether.
- the volume of the precipitation solvent and the alcohol solvent is preferably 1: 2 to 4: 1, more preferably 2: 1.
- the total amount of the precipitation solvent and the alcohol solvent is preferably such that at least the mixture of the RRRS, SSSR, RRSR and SSRS nebivolol intermediates is heated and dissolved, preferably the quality of the nebivolol intermediate mixture. 1 to 20 times.
- the heating temperature is preferably from 30 to 80 ° C, more preferably the solvent reflux temperature.
- the cooling temperature is preferably 0 to 50 ° C, more preferably 10 to 15 ° C.
- the time for depositing the crystals is preferably from 2 to 18 hours.
- the mixture of RRRS and SSSR type nebivolol intermediates of the formula II and formula III prepared by the present invention can be subjected to debenzylation to obtain nebivolol of the RRRS and SSSR type.
- the preparation method of the mixture of the RRRS, SSSR, RRSR and SSRS type nebivolol intermediates of the formula I can be obtained by the method of EP145067 and EP334429.
- Other reagents and materials used in the present invention are commercially available.
- the positive effect of the present invention is that the method of the present invention is simple to operate, has high separation efficiency, and is easily applied to industrial production. detailed description
- Example 2 The solid obtained in Example 2 was dissolved in 400 ml of ethanol, cooled to 40 ° C, concentrated hydrochloric acid was added thereto, and the pH was adjusted to 2-3, stirred for 1 hour, cooled, and crystallized. After incubation at 0 ° C for 6 hours, it was filtered, and the filter cake was washed once with a small amount of ethanol to give 68 g.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Plural Heterocyclic Compounds (AREA)
- Heterocyclic Carbon Compounds Containing A Hetero Ring Having Oxygen Or Sulfur (AREA)
Description
—种分离 RRRS和 SSSR^的奈必洛尔中间体混合物的方法 技术领域
本发明涉及一种化合物的异构体混合物的制备方法, 具体的涉及一种制备 RRRS和 SSSR型的奈必洛尔中间体混合物的方法。 背景技术
奈必洛尔如下式所示, 其结构中有 4个手性碳原子, 理论上有 16个立体异 构体, 考虑对称因素, 仅有 10个立体异构体存在。

其中, d-奈必洛尔 (RRRS )和 1-奈必洛尔 (SSSR) 为 β-受体阻滞剂, 具有 抗高血压作用。


a 5a


RRRS, SSSR, RRSR, SSRS
8a

RRRS, SSSR, RRSR, SSRS
在该专利中, 除 la、 2a、 3a、 4a和 5a外, 没有实施例具体描述制备过程, 也没有描述 7a和 7a的分离纯化过程,但可根据类似的方法制备得到。根据该专 利, 所得的为 (RRRS、 SSSR、 RRSR、 SSRS ) 奈必洛尔混合物。 US5759580 报道了采用分歩结晶的方法分离该混合物,但收率极低(6.6%)。 由此可以认为, 8a是制备奈必洛尔最关键的中间体,不仅因为 8a经脱苄基后即可得到奈必洛尔, 而且可通过控制 8a的立体构型直接得到想要的奈必洛尔立体异构体。 根据上述方法, 所得 8a为 (RRRS、 SSSR、 RRSR、 SSRS ) 的混合物:

(RRRS,SSSR,RRSR,SSRS) 其中 (RRRS , SSSR) 为所要的中间体:
WO2006016376报道将 8a转化成盐酸盐, 再分歩重结晶, 然后游离得到所 要的立体异构体, 该方法歩骤长, 收率低。
WO2007083318对此作了改进, 生成 8a后, 蒸干溶剂乙醇, 加入异丙醚, 所得固体再用异丙醚处理, 经两次处理后仍有 2%的其它异构体, 给其后的操作 带来不便。 发明内容
本发明所要解决的技术问题是为了克服现有的合成 RRRS和 SSSR型的奈必 洛尔的方法中, 分离 RRRS、 SSSR, RRSR和 SSRS型的中间体混合物的关键歩 骤操作不便且分离效率低的缺陷, 而提供一种简便而分离效率高的方法, 以制 备 RRRS和 SSSR型的奈必洛尔中间体混合物。
本发明的方法可为下述两种方式中的任一种:
( 1 ) 向含有如式 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中 间体混合物的醇类溶剂中, 加入沉淀溶剂, 加热, 冷却析出晶体, 过滤, 即可 制得如式 II和式 III分别所示的 RRRS和 SSSR型的奈必洛尔中间体的混合物。
(2 ) 将如式 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间体 混合物加入醇类溶剂和沉淀溶剂的混合溶剂中, 加热溶解, 然后冷却析出晶体, 过滤,即可制得如式 II和式 III分别所示的 RRRS和 SSSR型的奈必洛尔中间体的 混合物。

RRRS、 SSSR RRSR和 SSRS
 式 II
RRRS n
OH OH
.'、、、、' 式 III
SSSR
其中, X为氢, 或者 Cr^Qs的垸基或(^〜( 6的垸氧基, n为 1〜5; 较佳的 X为氢。
其中, 所述的醇类溶剂较佳的为甲醇、 乙醇、 丙醇和异丙醇中的一种或多 种, 更佳的为乙醇。 所述的沉淀溶剂较佳的为 C5〜C9烃类溶剂或醚类溶剂, 其 中包括环烃类, 较佳的为正已垸、 正庚垸和 /或环己垸, 更佳的为正已垸。 所述 的沉淀溶剂还可较佳的为醚类溶剂, 更佳的为异丙醚。 所述的沉淀溶剂和醇类 溶剂的体积比较佳的为 1 : 2〜4: 1, 更佳的为 2: 1。 所述的沉淀溶剂和醇类溶 剂的总量以至少使 RRRS、 SSSR, RRSR和 SSRS型的奈必洛尔中间体混合物加 热可溶解为宜, 较佳的为奈必洛尔中间体混合物质量的 1〜20倍。 所述的加热的 温度较佳的为 30〜80°C, 更佳的为溶剂回流温度。 所述的冷却的温度较佳的为 0〜50°C, 更佳的为 10〜15°C。 所述的析出晶体的时间较佳的为 2〜18小时。
本发明的一较佳实例中, 将采用乙醇为溶剂制得的含如式 IV所示的中间体 混合物 (X=氢) 的反应体系冷却至 50°C左右, 加入 2倍乙醇体积的正已垸, 加
热至回流, 然后冷却至 10〜15°C, 析晶 4小时, 过滤即分离得到如式 IV和式 V分 别所示的 RRRS和 SSSR型的奈必洛尔中间体的混合物。
本发明的另一较佳实例中, 将乙醇 /正己垸的体积比为 1 : 2的混合溶剂加入 如式 IV所示的中间体混合物 (X=氢) 中, 加热至回流, 然后冷却至 10〜15°C, 析晶 4小时,过滤即分离得到如式 V和式 VI分别所示的 RRRS和 SSSR型的奈必

RRRS、 SSSR RRSR和 SSRS
式 V

SSSR
本发明制得的如式 II和式 III分别所示的 RRRS和 SSSR型的奈必洛尔中间体 的混合物, 经脱苄后, 即可制得 RRRS和 SSSR型的奈必洛尔。

本发明的积极进歩效果在于: 本发明的方法操作简便 ΗΝ, 分离效率高, 易应 用于工业化生产。 具体实施方式
下面通过实施例的方式进一歩说明本发明, 但并不因此将本发明限制在所 述的实施例范围之中。
参考实施例 1 制备 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间体(式 I,
X=氢) 混合物
在 2000mL反应瓶中依次投入 1000 ml无水乙醇, 100 g(±)[lS*(R)]-6-氟 -3, 4-二氢 -2H-苯并吡喃 -2-环氧乙垸(参照 EP145067和 EP334429的方法制得, 7b) , 185g苄胺, 加热回流 6小时。 后冷却至 0°C, 保温 4小时过滤, 得滤饼, 40°C 烘干, 得 85g(±)[lS*(R)]-6-氟 -3, 4-二氢 -α-[[ (苯基甲基) 氨]甲基] -2Η-苯并吡
喃 -2-甲醇。
在 2000ml反应瓶中投入 500ml无水乙醇, 85g(±)[lS*(R)]-6-氟 -3, 4-二氢 - -[[ (苯基甲基) 氨]甲基] -2Η-苯并吡喃 -2-甲醇, 70g(±)[lS*(S)]-6-氟 -3, 4-二 氢 -2H-苯并吡喃 -2-环氧乙垸, 加热回流, 15小时后冷至 30°C, 制得含 RRRS、 SSSR、 RRSR和 SSRS型的 α α ' - [苯基甲基亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二 氢 -2Η-苯并吡喃 -2-甲醇] (式 IV混合物:)的乙醇溶液 ( 500ml )。
实施例 1 制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, X= H) 混合物
向参考实施例 1制得的溶液中, 加入 250ml正己垸, 加热至 30°C, 后冷却 至 10〜15°C, 保温结晶 4小时, 过滤, 得 67g (士) [R*[S*[S*(S*)]]] a α ' - [苯基甲 基亚氨基二 (亚甲基:)]双 -[6-氟 -3 , 4-二氢 -2Η-苯并吡喃 -2-甲醇] (式 V和式 VI的混合 物) (HPLC纯度 99.2%)。
实施例 2 制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, Χ= Η) 混合物
将参考实施例 1 制得的溶液减压回收乙醇得混合物 (式 R l60g, 加入 160g 乙醇: 异丙醚 =1 : 4的混合溶液, 加热至回流, 后冷却至 10〜15°C, 保温结晶 18 小时,过滤,得 68 g (士) [R*[S*[S*(S*)]]] a a ' - [苯基甲基亚氨基二 (亚甲基)]双 -[6- 氟 -3, 4-二氢 -2H-苯并吡喃 -2-甲醇] (式 V和式 VI的混合物) (HPLC纯度 99.6%)。 实施例 3制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, X= H) 混合物
向参考实施例 1制得的溶液中, 加入 1000ml正己垸, 加热至回流, 冷却至 10〜15°C, 保温结晶 4小时, 过滤, 得 67g (士) [R*[S*[S*(S*)]]] a α ' - [苯基甲基 亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二氢 -2Η-苯并吡喃 -2-甲醇] (式 V和式 VI的混合物)
(HPLC纯度 99.2%)。
实施例 4制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, X= H) 混合物
将参考实施例 1制得的溶液减压回收乙醇得混合物 (式 R l60g,加入 2318ml 乙醇: 正己垸 =1 : 2的混合溶液, 加热至回流, 然后冷却至 10〜15°C, 保温结晶 4小时, 过滤, 得 68 g (士) [R*[S*[S*(S*)]]] a a ' - [苯基甲基亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二氢 -2H-苯并吡喃 -2-甲醇] (式 V和式 VI的混合物) (HPLC 纯度 99.3% )o
参考实施例 2 制备 RRRS和 SSSR型的奈必洛尔
在 1000ml氢化反应釜中投入 300 ml无水乙醇, 67 g实施例 1制得的产物, 5gl0%钯碳, 通入氮气, 置换空气, 水浴加热到 50°C, 关闭氮气, 开始通氢气。 约 4小时, TLC检验, 反应结束。 滤去钯碳, 将滤液减压浓縮至滤液的 1/5时, 加入浓盐酸,测 PH值为 2-3,搅拌 1小时,冷却, 析晶。 0°C保温 6小时后过滤, 滤饼用少量乙醇洗涤一次, 得粗品 55g。 无水乙醇重结晶, 得 RRRS和 SSSR型 奈必洛尔 40g。
参考实施例 3 制备 RRRS和 SSSR型的盐酸奈必洛尔
实施例 2所得固体 66g加热溶于 400ml乙醇中, 冷至 40 °C, 加入浓盐酸, 测 pH值为 2-3, 搅拌 1小时, 冷却, 析晶。 0°C保温 6小时后过滤, 滤饼用少量 乙醇洗涤一次, 得 68g。
在 2000ml氢化反应釜中投入 1200 ml无水甲醇, 68 g上述产物, 5 gl0%钯 碳, 通入氮气, 置换空气, 水浴加热到 40°C, 关闭氮气, 开始通氢气。 约 4小 时, TLC 检验, 反应结束。 滤去钯碳, 将滤液减压浓縮至干, 无水乙醇重结晶 得 RRRS和 SSSR型的奈必洛尔 39g。
参考实施例 4制备 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间体(式 I,
X=4-甲氧基, n=l ) 混合物
在100011^反应瓶中依次投入500 1111乙醇, 100 g(±)[lS*(R)]-6-氟 -3, 4-二 氢 -2H-苯并吡喃 -2-环氧乙垸(参照 EP145067和 EP334429的方法制得, 7b), 170 g 4-甲氧基苄胺, 加热回流 24小时。 后冷却至 0°C, 保温 4小时过滤, 得滤饼, 40°C烘干, 得 94g(±)[lS*(R)]-6-氟 -3, 4-二氢 -α-[[ (4-甲氧基苯基甲基) 氨]甲 基] -2Η-苯并吡喃 -2-甲醇。
在 2000ml反应瓶中投入 300ml甲醇, 94g(±)[lS*(R)]-6-氟 -3, 4-二氢 -α-[[(4- 甲氧基苯基甲基) 氨]甲基] -2Η-苯并吡喃 -2-甲醇, 67 g(±)[lS*(S)]-6-氟 -3, 4-二 氢 -2H-苯并吡喃 -2-环氧乙垸, 加热回流, 15小时后冷至 30°C, 制得含 RRRS、 SSSR, RRSR和 SSRS型的 α α ' -[4-甲氧基苯基甲基亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二氢 -2Η-苯并吡喃 -2-甲醇] (式 IV混合物)的乙醇溶液 (300ml)。
实施例 5制备 RRRS和 SSSR型的奈必洛尔中间体(式 I, X= 4-甲氧基)混合 物
向参考实施例 4制得的溶液中, 加入 900ml正庚垸, 加热至 50°C, 冷却至 0°C, 保温结晶 2小时, 过滤, 得 70 g RRRS和 SSSR型的奈必洛尔中间体 (式 I, X=4-甲氧基) 混合物 (HPLC纯度 99.6%)。
实施例 6制备 RRRS和 SSSR型的奈必洛尔中间体(式 I, X=4-甲氧基)混合 物
将参考实施例 4制得的溶液减压回收乙醇得固体混合物( 162g),加入 3240g 乙醇: 环己垸 =1 : 3 的混合溶液, 加热至 80°C, 后冷却至 50°C, 保温结晶 18 小时, 过滤, 得 77 g RRRS和 SSSR型的奈必洛尔中间体 (式 I, X=4-甲氧基) 混合物 (HPLC纯度 99.2%)。
参考实施例 5制备 RRRS、SSSR、RRSR和 SSRS型的奈必洛尔中间体(式
I, X=4-甲基, n=l )混合物
在100011^反应瓶中依次投入500 1111异丙醇, 100 g(±)[lS*(R)]-6-氟 -3, 4- 二氢 -2H-苯并吡喃 -2-环氧乙垸, 170 g4-甲基苄胺, 加热回流 6小时。 后冷却至 0°C, 保温 4小时过滤, 得滤饼 97 g(±)[lS*(R)]-6-氟 -3, 4-二氢 -α-[[ (4-甲基苯 基甲基) 氨]甲基] -2Η-苯并吡喃 -2-甲醇。
在 2000ml 反应瓶中投入 300ml 异丙醇, 97g(±)[lS*(R)]-6-氟 -3, 4-二氢 -α-[[ (4-甲基苯基甲基)氨]甲基] -2H-苯并吡喃 -2-甲醇, 67.5g(±)[lS*(S)]-6-氟 -3, 4-二氢 -2H-苯并吡喃 -2-环氧乙垸,,加热回流, 15小时后冷至 50°C,制得含 RRRS、 SSSR、RRSR和 SSRS型的 α α ' -[4-甲基苯基甲基亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二氢 -2Η-苯并吡喃 -2-甲醇]异丙醇溶液 (300ml)。
实施例 7制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, X=4-甲基, n=l ) 混合物
向参考实施例 5制得的溶液中, 加入 600ml石油醚, 加热至 70°C, 冷却至 15°C, 保温结晶 10小时, 过滤, 得 75 g含 RRRS、 SSSR型的 α α ' -[4-甲基苯 基甲基亚氨基二 (亚甲基)]双 -[6-氟 -3, 4-二氢 -2Η-苯并吡喃 -2-甲醇] (HPLC纯度 99.1
实施例 8制备 RRRS和 SSSR型的奈必洛尔中间体 (式 I, X=4-甲基, n=l ) 混合物
将参考实施例 5制得的溶液减压回收乙醇得混合物 (163g), 中加入 1630g 乙醇: 正壬垸 =1 : 1 的混合溶液, 加热至回流, 后冷却至 10°C, 保温结晶 4小 时, 过滤, 得 76 g含 RRRS、 SSSR型的 α α ' -[4-甲基苯基甲基亚氨基二 (亚甲 基)]双 -[6-氟 -3, 4-二氢 -2Η-苯并吡喃 -2-甲醇] (HPLC纯度 99.3%)。
Claims
1. 一种制备如式 II和式 III分别所示的 RRRS和 SSSR型的奈必洛尔中间体的混 合物的方法, 其特征在于任选下述两种方式中的一种:
( 1 ) 向含有如式 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间 体混合物的醇类溶剂中, 加入沉淀溶剂, 加热溶解, 冷却析出晶体, 过滤, 即可;
(2 )将如式 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间体混 合物加入醇类溶剂和沉淀溶剂的混合溶剂中,加热溶解,然后冷却析出晶体, 过滤, 即可;

RRRS
 式 III
SSSR
其中, X为 H, 或者 Cr^ 的垸基或(^〜03的垸氧基, n为 1〜5。
2. 如权利要求 1所述的方法, 其特征在于: 所述的醇类溶剂为甲醇、 乙醇、 丙 醇和异丙醇中的一种或多种。
3. 如权利要求 2所述的方法, 其特征在于: 所述的醇类溶剂为乙醇。
4. 如权利要求 1所述的方法, 其特征在于:所述的沉淀溶剂为烃类溶剂和 /或醚
5. 如权利要求 4所述的方法,其特征在于:所述的烃类溶剂为 C5〜C9烃类溶剂; 所述的醚类溶剂为异丙醚。
6. 如权利要求 5所述的方法, 其特征在于: 所述的 C5〜C9烃类溶剂为正已垸、 正庚垸、 石油醚和环己垸中的一种或多种。
7. 如权利要求 6所述的方法, 其特征在于: 所述的 C5〜C9烃类溶剂为正已垸。
8. 如权利要求 1所述的方法, 其特征在于: 所述的沉淀溶剂和醇类溶剂的体积 比为 1 : 2〜4: 1。
9. 如权利要求 8所述的方法, 其特征在于: 所述的沉淀溶剂和醇类溶剂的体积 比为 2: 1。
10.如权利要求 1所述的方法, 其特征在于: 所述的加热的温度为 30〜80°C。
11.如权利要求 10所述的方法, 其特征在于: 所述的加热的温度为溶剂回流温
12.如权利要求 1所述的方法, 其特征在于: 所述的冷却的温度为 0〜50°C。
13.如权利要求 12所述的方法, 其特征在于: 所述的冷却的温度为 10〜15°C。
14.如权利要求 1所述的方法, 其特征在于: 所述的析出晶体的时间为 2〜18小 时。
15.如权利要求 1所述的方法, 其特征在于: 向含如式 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间体混合物的乙醇溶液中, 加入 2倍乙醇体 积的正已垸, 加热至回流, 然后冷却至 10〜15°C, 析晶 4小时, 过滤即可。
16.如权利要求 1所述的方法, 其特征在于: 将体积比为 1 : 2的乙醇和正己垸 的混合溶剂加入 I所示的 RRRS、 SSSR、 RRSR和 SSRS型的奈必洛尔中间 体混合物中, 加热至回流, 然后冷却至 10〜15°C, 析晶 4小时, 过滤即可。
Priority Applications (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| EP08867602.8A EP2236510B1 (en) | 2007-12-21 | 2008-12-12 | Process for isolation of a mixture of rrrs and sssr configurations of nebivolol intermediates |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| CN200710172796.9 | 2007-12-21 | ||
| CN2007101727969A CN101463024B (zh) | 2007-12-21 | 2007-12-21 | 一种制备rrrs和sssr型的奈必洛尔中间体混合物的方法 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2009082913A1 true WO2009082913A1 (en) | 2009-07-09 |
Family
ID=40803844
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/CN2008/073492 Ceased WO2009082913A1 (en) | 2007-12-21 | 2008-12-12 | Process for isolation of a mixture of rrrs and sssr configurations of nebivolol intermediates |
Country Status (3)
| Country | Link |
|---|---|
| EP (1) | EP2236510B1 (zh) |
| CN (1) | CN101463024B (zh) |
| WO (1) | WO2009082913A1 (zh) |
Cited By (4)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP2907810A1 (en) | 2014-02-14 | 2015-08-19 | Corden Pharma International GmbH | A new method for producing nebivolol hydrochloride of high purity |
| EP2907809A1 (en) | 2014-02-14 | 2015-08-19 | Corden Pharma International GmbH | Base-free process for the preparation of ketone intermediates usable for manufacture of nebivolol |
| DE102014107132A1 (de) | 2014-05-20 | 2015-11-26 | Corden Pharma International Gmbh | Verfahren zur Herstellung von Epoxiden die in der Herstellung von Nebivolol und dessen Derivaten einsetzbar sind |
| US10526304B2 (en) | 2015-05-19 | 2020-01-07 | Zhejiang Ausun Pharmaceutical Co., Ltd. | Nebivolol synthesis method and intermediate compound thereof |
Families Citing this family (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN105085499B (zh) * | 2015-08-07 | 2018-09-25 | 上海现代制药海门有限公司 | 盐酸奈必洛尔中间体混合物的结晶分离方法 |
Citations (5)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0145067A2 (en) | 1983-12-05 | 1985-06-19 | Janssen Pharmaceutica N.V. | Derivatives of 2,2'-iminobisethanol |
| EP0334429A1 (en) | 1988-03-23 | 1989-09-27 | Janssen Pharmaceutica N.V. | Agents for lowering the blood pressure |
| US5759580A (en) | 1994-02-17 | 1998-06-02 | Janssen Pharmaceutica, N.V. | Compositions containing micronized nebivolol |
| WO2006016376A1 (en) | 2004-08-11 | 2006-02-16 | Hetero Drugs Limited | A novel process for preparation of nebivolol intermediates |
| WO2007083318A1 (en) | 2006-01-18 | 2007-07-26 | Hetero Drugs Limited | Process for isolation of desired isomers of nebivolol intermediates |
-
2007
- 2007-12-21 CN CN2007101727969A patent/CN101463024B/zh active Active
-
2008
- 2008-12-12 EP EP08867602.8A patent/EP2236510B1/en active Active
- 2008-12-12 WO PCT/CN2008/073492 patent/WO2009082913A1/zh not_active Ceased
Patent Citations (5)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0145067A2 (en) | 1983-12-05 | 1985-06-19 | Janssen Pharmaceutica N.V. | Derivatives of 2,2'-iminobisethanol |
| EP0334429A1 (en) | 1988-03-23 | 1989-09-27 | Janssen Pharmaceutica N.V. | Agents for lowering the blood pressure |
| US5759580A (en) | 1994-02-17 | 1998-06-02 | Janssen Pharmaceutica, N.V. | Compositions containing micronized nebivolol |
| WO2006016376A1 (en) | 2004-08-11 | 2006-02-16 | Hetero Drugs Limited | A novel process for preparation of nebivolol intermediates |
| WO2007083318A1 (en) | 2006-01-18 | 2007-07-26 | Hetero Drugs Limited | Process for isolation of desired isomers of nebivolol intermediates |
Non-Patent Citations (1)
| Title |
|---|
| See also references of EP2236510A4 * |
Cited By (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP2907810A1 (en) | 2014-02-14 | 2015-08-19 | Corden Pharma International GmbH | A new method for producing nebivolol hydrochloride of high purity |
| EP2907809A1 (en) | 2014-02-14 | 2015-08-19 | Corden Pharma International GmbH | Base-free process for the preparation of ketone intermediates usable for manufacture of nebivolol |
| DE102014107132A1 (de) | 2014-05-20 | 2015-11-26 | Corden Pharma International Gmbh | Verfahren zur Herstellung von Epoxiden die in der Herstellung von Nebivolol und dessen Derivaten einsetzbar sind |
| US10526304B2 (en) | 2015-05-19 | 2020-01-07 | Zhejiang Ausun Pharmaceutical Co., Ltd. | Nebivolol synthesis method and intermediate compound thereof |
| US11142512B2 (en) | 2015-05-19 | 2021-10-12 | Zhejiang Ausun Pharmaceutical Co., Ltd. | Nebivolol synthesis method and intermediate compound thereof |
| US12466802B2 (en) | 2015-05-19 | 2025-11-11 | Zhejiang Ausun Pharmaceutical Co., Ltd. | Nebivolol synthesis method and intermediate compound thereof |
Also Published As
| Publication number | Publication date |
|---|---|
| EP2236510A1 (en) | 2010-10-06 |
| CN101463024B (zh) | 2011-06-08 |
| CN101463024A (zh) | 2009-06-24 |
| EP2236510A4 (en) | 2011-11-16 |
| EP2236510B1 (en) | 2014-04-30 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| KR101377778B1 (ko) | 엘트롬보팩 및 엘트롬보팩 염의 다형체 및 이의 제조 방법 | |
| JP2839344B2 (ja) | 環状アミノ酸、並びにその中間体の製造方法 | |
| IL149050A (en) | Process for preparation of sertraline hyrochloride polymorphic form ii | |
| JP2008239629A (ja) | 新規なベンラファクシン塩酸塩多型形状、ならびにその調製方法 | |
| JP7097467B2 (ja) | ブリバラセタム中間体、その製造方法及びブリバラセタムの製造方法 | |
| CZ302424B6 (cs) | Zpusob prípravy almotriptanu o vysoké cistote | |
| WO2015111085A2 (en) | Processes for the preparation of eltrombopag and pharmaceutically acceptable salts, solvates and intermediates thereof | |
| TWI491607B (zh) | 4,4’-(1-甲基-1,2-乙二基)-雙-(2,6-哌二酮)之新製法 | |
| CN101463024B (zh) | 一种制备rrrs和sssr型的奈必洛尔中间体混合物的方法 | |
| EP4046687A1 (en) | Method for producing centanafadine | |
| WO2010046360A1 (en) | Process for the preparation of substantially optically pure repaglinide and precursors thereof | |
| CN118420493B (zh) | 一种2,5-二取代氨基己酸酯的制备方法及使用其制备利特昔替尼的方法 | |
| CN114890999B (zh) | 一种pqq的制备方法 | |
| TW202019872A (zh) | 稠合三環γ-胺基酸衍生物的製備方法及其中間體 | |
| CN113072514A (zh) | 轮环藤宁及其中间体的制备方法 | |
| EP3188737A1 (en) | Crystalline form of eltrombopag free acid | |
| WO2004026860A1 (ja) | 結晶析出方法 | |
| US20090076288A1 (en) | Process for isolation of desired isomers of nebivolol intermediates | |
| CN111100121B (zh) | 黄连素或其盐酸盐的纯化方法 | |
| CN103467350A (zh) | (s)-氮杂环丁烷-2-羧酸的制备方法 | |
| CN109280011B (zh) | Oled中间体2-溴芘的合成法 | |
| CN113024433A (zh) | 一种氨磺必利亚砜杂质的制备方法 | |
| US5516937A (en) | Process for preparing aminoethylglycine | |
| CN115872873B (zh) | 一种双(2-溴乙基)胺氢溴酸盐的重结晶纯化方法 | |
| EP1539751A1 (en) | Process for the preparation of imidazo(1,2-a)pyridine-3-acetamides |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 08867602 Country of ref document: EP Kind code of ref document: A1 |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2008867602 Country of ref document: EP |