WO2005016881A1 - Bicyclische indolinsulfonamid-derivate - Google Patents

Bicyclische indolinsulfonamid-derivate Download PDF

Info

Publication number
WO2005016881A1
WO2005016881A1 PCT/EP2004/008075 EP2004008075W WO2005016881A1 WO 2005016881 A1 WO2005016881 A1 WO 2005016881A1 EP 2004008075 W EP2004008075 W EP 2004008075W WO 2005016881 A1 WO2005016881 A1 WO 2005016881A1
Authority
WO
WIPO (PCT)
Prior art keywords
formula
compounds
methyl
meanings given
hydrogen
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/EP2004/008075
Other languages
English (en)
French (fr)
Inventor
Elke Dittrich-Wengenroth
Hilmar Bischoff
Stephan Siegel
Michael Woltering
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
Bayer AG
Original Assignee
Bayer Healthcare AG
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Application filed by Bayer Healthcare AG filed Critical Bayer Healthcare AG
Priority to EP04763344A priority Critical patent/EP1654227A1/de
Priority to CA002534171A priority patent/CA2534171A1/en
Priority to US10/566,340 priority patent/US20070117860A1/en
Priority to JP2006522267A priority patent/JP2007501195A/ja
Publication of WO2005016881A1 publication Critical patent/WO2005016881A1/de
Anticipated expiration legal-status Critical
Ceased legal-status Critical Current

Links

Classifications

    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07D—HETEROCYCLIC COMPOUNDS
    • C07D209/00—Heterocyclic compounds containing five-membered rings, condensed with other rings, with one nitrogen atom as the only ring hetero atom
    • C07D209/02—Heterocyclic compounds containing five-membered rings, condensed with other rings, with one nitrogen atom as the only ring hetero atom condensed with one carbocyclic ring
    • C07D209/04—Indoles; Hydrogenated indoles
    • C07D209/08—Indoles; Hydrogenated indoles with only hydrogen atoms or radicals containing only hydrogen and carbon atoms, directly attached to carbon atoms of the hetero ring
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P3/00—Drugs for disorders of the metabolism
    • A61P3/04—Anorexiants; Antiobesity agents
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00—Drugs for disorders of the cardiovascular system
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00—Drugs for disorders of the cardiovascular system
    • A61P9/10—Drugs for disorders of the cardiovascular system for treating ischaemic or atherosclerotic diseases, e.g. antianginal drugs, coronary vasodilators, drugs for myocardial infarction, retinopathy, cerebrovascula insufficiency, renal arteriosclerosis

Definitions

  • the present application relates to new bicyclic indoline sulfonamide derivatives, processes for their preparation and their use in medicaments, in particular as potent PPAR-delta activating compounds for the prophylaxis and or treatment of cardiovascular diseases, in particular dyslipidemias, arteriosclerosis and coronary heart diseases.
  • fibrates are the only form of therapy for patients in these risk groups. They act as weak agonists of the peroxisome proliferator-activated receptor (PPAR) -alpha (Nature 1990, 347, 645-50). A disadvantage of previously approved fibrates is their poor interaction with the receptor, which leads to high daily doses and significant side effects.
  • PPAR peroxisome proliferator-activated receptor
  • WO 00/23407 discloses PPAR modulators for the treatment of obesity, atherosclerosis and / or diabetes.
  • WO 93/15051 and EP 636 608-A1 describe 1-benzenesulfonyl-1,3-dihydroindol-2-one derivatives as vasopressin and / or oxytocin antagonists for the treatment of various diseases.
  • the object of the present invention was to provide new compounds which can be used as PPAR delta modulators.
  • the present invention relates to compounds of the general formula (I)
  • R 1 represents phenyl or 5- to 6-membered heteroaryl with up to two heteroatoms from the series N, O and or S, which in turn are each one to three times, identical or different, by substituents selected from the group halogen, cyano , Nitro, (CC 6 ) - alkyl, which in turn can be substituted by hydroxy, (-C-C 6 ) alkoxy, trifluoromethyl, trifluoromethoxy, (C ⁇ -C 6 ) alkylsulfonyb (CC 6 ) alkanoyb (CC 6 ) -Alkoxy-carbonyb-carboxyb amino, (-C-C 6 ) -acylamino, mono- and di- (-C-C 6 ) -alkylamino may be substituted,
  • R 2 and R 3 are the same or different and independently of one another represent hydrogen or (Cj-C) - alkyl or together with the carbon atom to which they are attached form a 3- to 7-membered, spiro-linked cycloalkyl ring .
  • R 4 represents hydrogen or (C, -C 4 ) -alkyl
  • R s and R 6 are the same or different and independently of one another represent hydrogen or (-CC 4 ) - alkyl
  • R 7 represents hydrogen or a hydrolyzable group which can be broken down into the corresponding carboxylic acid
  • n the number 1 or 2
  • groups are exemplary and preferably: benzyb (CC 6 ) alkyl or (C 3 -C 8 ) cycloalkyb, each optionally one or more times, identically or differently, by halogen, hydroxyl, amino, (CC 6 ) alkoxy, Carboxyb (CC 6 ) alkoxycarbonyb (-C-C 6 ) alkoxycarbonylamino or (C ⁇ -C 6 ) alkanoyloxy are substituted, or in particular (C r C 4 ) -alky which may be one or more, identical or different, by halogen , Hydroxy, amino, (CC 4 ) alkoxy, carboxyb (CC 4 ) alkoxycarbonyl (CC) alkoxycarbonylamino or (C]
  • C 1 -C 6 -alkyl and (CC 4 ) -alkyl stand for a straight-chain or branched alkyl radical having 1 to 6 or 1 to 4 carbon atoms.
  • the following may be mentioned as examples and preferably: methyl, ethyl n-propyb isopropyl and tert-butyb
  • (C Cg) cycloalkyl stands for a monocyclic cycloalkyl group with 3 to 8 carbon atoms. Examples and preferably mentioned are: Cyclopropyb Cyclobutyb Cyclopentyl and Cyclohexyb
  • (Cr-) alkoxy and (Cv-CHAlkoxY) stand for a straight-chain or branched alkoxy radical with 1 to 6 or 1 to 4 carbon atoms.
  • a straight-chain or branched alkoxy radical with 1 to 4 carbon atoms is preferred called: methoxy, ethoxy, n-propoxy, isopropoxy and tert-butoxy.
  • (C 1 -Cg) -alkoxycarbonyl and (-G -alkoxycarbonyl) represent a straight-chain or branched alkoxy radical having 1 to 6 or 1 to 4 carbon atoms, which is linked via a carbonyl group.
  • a straight-chain or branched alkoxycarbonyl radical is preferred with 1 to 4 carbon atoms, and examples which may be mentioned are: methoxycarbonyl, ethoxycarbonyl, n-propoxycarbonyl, isopropoxycarbonyl and tert-butoxycarbonyl
  • (-CfiVAlkoxycarbonylamino and (CC ⁇ -alkoxycarbonylamino) stand in the context of the invention for an amino group with a straight-chain or branched alkoxycarbonyl substituent which has 1 to 6 or 1 to 4 carbon atoms in the alkoxy radical and is linked via the carbonyl group.
  • Preferred an alkoxycarbonylamino radical having 1 to 4 carbon atoms, and examples which may be mentioned are: methoxycarbonylamino, ethoxycarbonylamino, n-propoxycarbonylamino and tert-butoxycarbonylamino.
  • fCq-Cg) -alkanoyl represents a straight-chain or branched alkyl radical having 1 to 6 carbon atoms, which carries a double-bonded oxygen atom in the 1 position and is linked via the 1 position.
  • a straight-chain or branched alkanoyl radical having 1 to 4 carbon atoms is preferred. Examples and preferably mentioned are: Formyb Acetyb Propionyb n-Butyryb i-Butyryb Pivaloyl and n-Hexanoyb
  • (Cr-CsValkanoyloxy and (Cr-C balkanoyloxy) stand for a straight-chain or branched alkyl radical having 1 to 6 or 1 to 4 carbon atoms, which carries a double-bonded oxygen atom in the 1 position and in which Position is linked via a further oxygen atom.
  • An alkanoyloxy radical having 1 to 4 carbon atoms is preferred. Examples and preferably mentioned are: acetoxy, propionoxy, n-butyroxy, i-butyroxy, pivaloyloxy, n-hexanoyloxy.
  • Mono- (C 1 -C ⁇ -alkylamino and mono- (C 1 -C 4) alkylamino are in the context of the invention an amino group having a straight-chain or branched alkyl substituent which has 1 to 6 or 1 to 4 carbon atoms.
  • a straight-chain or branched monoalkylamino radical having 1 to 4 carbon atoms is preferred, and examples which may be mentioned are: methylamino, ethylamino, n-propylamino, isopropylamino and tert-butylamino.
  • Di- (C 1 -C ⁇ ) -All / lamino and DIJG-CaVAlkylamino are in the context of the invention an amino group having two identical or different straight-chain or branched alkyl substituents having in each case 1 to 6 or 1 to 4 carbon atoms.
  • Straight-chain or 'branched dialkylamino radicals are preferably each having 1 to 4 carbon atoms.
  • Examples that may be mentioned are: NN-dimethylamino, N / V-diethylamino, H-ethyl-H-methylamino, N-methyl-H-n-propylamino, TV-isopropyl-Nn-propylamino, N-tert-butyl -H-memylamino, N-ethyl-Nn-pentylamino and Nn- hexyl-H-memylanjJ.no.
  • (C 1 -C 6 -acylamino in the context of the invention represents an amino group with a straight-chain or branched alkanoyl substituent which has 1 to 6 carbon atoms and is linked via the carbonyl group.
  • An acylamino radical with 1 to 2 carbon atoms for example and preferably the following are mentioned: formamido, acetamido, propionamido, n-butyramido and pivaloylamido.
  • (C Cfi) alkylsulfonyl represents a straight-chain or branched alkylsulfonyl radical having 1 to 6 carbon atoms.
  • a straight-chain or branched alkylsulfonyl radical having 1 to 4 carbon atoms is preferred.
  • the following may be mentioned as examples and preferably: methylsulfonyb ethylsulfonyb n-propylsulfonyb isopropylsulfonyb tert.-butylsulfonyb n-pentylsulfonyl and n-hexylsulfonyb
  • 5- to 6-membered heteroaryl with up to 2 identical or different heteroatoms from the series N, O and / or S in the context of the invention represents a monocyclic aromatic heterocycle (heteroaromatics) which has a ring carbon atom or optionally a ring nitrogen atom of the heteroaromatic is linked.
  • Halogen in the context of the invention includes fluorine, chlorine, bromine and iodine. Chlorine or fluorine are preferred.
  • the compounds according to the invention can exist in stereoisomeric forms which either behave like image and mirror image (enantiomers) or do not behave like image and mirror image (diastereomers).
  • the invention relates both to the enantiomers or diastereomers and to their respective mixtures. Like the diastereomers, the racemic forms can be separated into the stereoisomerically uniform constituents in a known manner.
  • the compounds according to the invention can also be present as salts.
  • physiologically acceptable salts are preferred.
  • Physiologically acceptable salts can be salts of the compounds according to the invention with inorganic or organic acids.
  • Salts with inorganic acids such as, for example, hydrochloric acid, hydrobromic acid, phosphoric acid or sulfuric acid, or salts with organic carboxylic or sulfonic acids such as, for example, acetic acid, propionic acid, maleic acid, fumaric acid, malic acid, citric acid, tartaric acid, lactic acid, benzoic acid, or methanesulfonic acid, ethanesulfonic acid , Benzenesulfonic acid, toluenesulfonic acid or naphthalenedisulfonic acid.
  • Physiologically acceptable salts can also be salts of the compounds according to the invention with bases, such as metal or ammonium salts.
  • bases such as metal or ammonium salts.
  • alkali metal salts for example sodium or potassium salts
  • alkaline earth metal salts for example magnesium or calcium salts
  • ammonium salts which are derived from ammonia or organic amines, for example ethylamine, di- or triethylamine, ethyldiisopropylamine, monoethanolamine , Di- or triethanolamine, dicyclohexylamine, dimethylaminoethanob dibenzylamine, N-methylmorpholine, dihydroabietylamine, 1-ephenamine, methylpiperidine, arginine, lysine, ethylenediamine or 2-phenylethylamine.
  • the compounds according to the invention can also be in the form of their solvates, in particular in the form of their hydrates.
  • R 1 represents phenyl, the one or two, identical or different, by substituents selected from the group fluorine, chlorine, cyano, (C r C 4 ) alkylb (C 1 -C 4 ) alkoxy, trifluoromethyl Trifluoromethoxy, methylsulfonyb ' .
  • Acetyb Propionyb (-C-C 4 ) -alkoxycarbonyl amino, acetylamino, mono- and di- (-C-C 4 ) -alkylamino can be substituted
  • R 2 and R 3 are the same or different and independently of one another represent hydrogen or (-C) alkyl or together with the carbon atom to which they are attached form a 5- to 6-membered, spiro-linked cycloalkyl ring,
  • R 4 represents hydrogen or methyl
  • R 5 and R 6 are the same or different and are independently hydrogen or methyl
  • R 7 represents hydrogen
  • n stands for the number 1 or 2.
  • R 1 represents phenyl, which can be substituted one to two times, identically or differently, by substituents selected from the group consisting of fluorine, chlorine, methyl, trifluoromethyl and trifluoromethoxy,
  • R 2 represents methyl
  • R 3 represents methyl
  • R 2 and R 3 together with the carbon atom to which they are attached form a spiro-linked cyclopentane or cyclohexane ring,
  • R 4 represents hydrogen or methyl
  • R 5 and R 6 each represent hydrogen
  • R 7 represents hydrogen
  • n stands for the number 1 or 2.
  • radical definitions given above apply both to the end products of the formula (I) and correspondingly to the starting materials or intermediates required in each case for the preparation.
  • radical definitions specified in detail in the respective combinations or preferred combinations of radicals are also replaced as desired by radical definitions of other combinations, irrespective of the respectively specified combinations of the radicals.
  • R represents phenyl which is substituted by fluorine, chlorine or trifluoromethyl
  • n stands for the number 1 or 2.
  • Y represents chlorine or bromine
  • PG stands for a suitable amino protective group, preferably for 4-nitrophenylsulfonyl
  • R 8 represents hydrogen or methyl or both radicals together form a CH 2 CH 2 or C (CH 3 ) 2 -C (CH 3 ) 2 bridge,
  • T represents benzyl or (CC 6 ) alkyl
  • these carboxylic acids (LX) are further modified to give compounds of the formula (I) by known esterification methods, and the resulting compounds of the formula (LX) or (I) are optionally reacted with the corresponding (i) solvents and / or (ii) bases or acids to give their solvates, salts and / or solvates of the salts.
  • Inert solvents for process step (ITJ) + (IV) - »(V) are, for example, ethers such as diethyl ether, dioxane, tetrahydrofuran, glycol dimethyl ether or diethylene glycol dimethyl ether, alcohols such as methanol, ethanol, n-propanol, iso-propanob n-butanol or tert.
  • ethers such as diethyl ether, dioxane, tetrahydrofuran, glycol dimethyl ether or diethylene glycol dimethyl ether
  • alcohols such as methanol, ethanol, n-propanol, iso-propanob n-butanol or tert.
  • -Butanob hydrocarbons such as benzene, xylob toluob hexane, cyclohexane or petroleum fractions, or other solvents such as dimethylformamide, acetonitrile
  • the usual inorganic or organic bases are suitable as bases for the process step (HI) + (IV) - »(V).
  • bases preferably include alkali metal hydroxides such as lithium, sodium or potassium hydroxide, alkali metal or alkaline earth metal carbonates such as sodium, potassium or calcium carbonate, alkali metal phosphates such as sodium or potassium phosphate, or organic amines such as pyridine, triethylamine, ethyldiisopropylamine, N-methylmorpholine or N-methylpiperidine.
  • Sodium or potassium carbonate or potassium phosphate are particularly preferred.
  • the base is used in an amount of 1 to 5, preferably 2 to 3 mob based on 1 mol of the compound of formula (TU).
  • Suitable palladium catalysts for process step (ITJ) + (TV) - »(V) are preferably palladium (O) - or palladium (ITj compounds which are used preformed, such as, for example, [1,1'-bis (diphenylphosphino) ferrocenyl] palladium ( ⁇ ) chloride, bis (triphenylphosphine) - palladium ( ⁇ ) chloride or tetrakis (triphenylphosphine) palladium (0), or those obtained in situ from a suitable palladium source such as bis (dibenzylidene acetone) palladium (0) and a suitable phosphine ligand can be generated.
  • a suitable palladium source such as bis (dibenzylidene acetone) palladium (0) and a suitable phosphine ligand can be generated.
  • the reaction generally takes place in a temperature range from 0 ° C. to + 150 ° C., preferably from + 20 ° C. to + 120 ° C.
  • the reaction can be carried out at normal, elevated or reduced pressure (e.g. from 0.5 to 5 bar). Generally one works at normal pressure.
  • Inert solvents for process step (VI) + (VU) -> (VI-LT) are, for example, halogenated hydrocarbons such as dichloromethane, trichloromethane, carbon tetrachloride, trichloroethane, tetrachloroethane, 1,2-dichloroethane or trichlorethylene, ethers such as diethyl ether, dioxane, tetrahydrofuran , Glycol dimethyl ether or diethylene glycol dimethyl ether, hydrocarbons such as benzene, xylobotoluene hexane, cyclohexane or petroleum fractions, or other solvents such as nitro methane, ethyl acetate, acetone, dimethylformamide, dimethyl sulfoxide, acetonitrile, N-methylpyrrolidinone or pyridine. It is also possible to use mixtures of the solvents mentioned.
  • the usual inorganic or organic bases are suitable as bases for process step (VI) + (VII) - »(VITJ).
  • bases preferably include alkali metal hydroxides such as lithium, sodium or potassium hydroxide, alkali metal or alkaline earth metal carbonates such as sodium, potassium or calcium carbonate, alkali metal hydrides such as sodium hydride, or organic amines such as pyridine, triethylamine, ethyldiisopropylamine, N-methylmorpholine or N methylpiperidine.
  • Amine bases such as triethylamine, pyridine or ethyldiisopropylamine are particularly preferred, optionally in the presence of catalytic amounts (approx. 10 mol%) of 4-NN-dimethylaminopyridine or 4-pyrrolidinopyridine.
  • the base is used here in an amount of 1 to 5, preferably 1 to 2.5 mob based on 1 mol of the compound of the formula (VIT).
  • the reaction generally takes place in a temperature range from -20 ° C to + 100 ° C, preferably from 0 ° C to +75 ° C.
  • the reaction can be normal, increased or reduced
  • Pressure e.g. from 0.5 to 5 bar. Generally one works at normal pressure.
  • Inert solvents for process step (VITI) - »(LX) are, for example, halogenated hydrocarbons such as dichloromethane, 1,2-dichloroethane or trichlorethylene, ethers such as diethyl ether, dioxane, tetrahydrofuran, glycol dimethyl ether or diethylene glycol dimethyl ether, alcohols such as methanol, ethanob n-propanob isopropane n-Butanol or tert-butanob hydrocarbons such as benzene, xylobotoluene hexane, cyclohexane or petroleum fractions, or other solvents such as nitromethane, acetone, dimethylformamide, dimethyl sulfoxide, acetonitrib N-methylpyrrolidine or water. It is also possible to use mixtures of the solvents mentioned. Alcohols such as methanol or ethanol and mixtures thereof with tetra
  • the usual inorganic bases are suitable as bases for process step (VITJ) - (LX). These preferably include alkali hydroxides such as lithium, sodium or potassium hydroxide, or alkali or alkaline earth carbonates such as sodium, potassium or calcium carbonate. Lithium or sodium hydroxide are particularly preferred.
  • the base is used in an amount of 1 to 5, preferably 1 to 3 mob based on 1 mol of the compound of formula (VWL).
  • Suitable acids for process step (NIJJ) - »(LX) are the customary inorganic acids such as, for example, hydrochloric acid or sulfuric acid, or sulfonic acids such as toluenesulfonic acid, methanesulfonic acid or trifluoromethanesulfonic acid, or carboxylic acids such as trifluoroacetic acid.
  • the reaction generally takes place in a temperature range from -20 ° C to + 100 ° C, preferably from 0 ° C to + 30 ° C.
  • the reaction can be normal, increased or reduced
  • Pressure e.g. from 0.5 to 5 bar. Generally one works at normal pressure.
  • the compounds of the formula (11) are known or can be prepared analogously to processes known from the literature, for example by
  • Inert solvents for process step (X) + (XI) ⁇ (XU) or (XITJ) are, for example, halogenated hydrocarbons such as dichloromethane, trichloromethane, tetrachloromethane, trichloroethane, tetrachloroethane, 1,2-dichloroethane or trichlorethylene, ethers such as dioxane, tetrahydrofuran, Glycol dimethyl ether or diethylene glycol dimethyl ether, alcohols such as methanol, ethanob n-propanob iso-propanob n-butanol or tert-butanob or hydrocarbons such as benzene, xylob toluene hexane, cyclohexane or petroleum fractions, or other solvents such as acetonitrile or water.
  • halogenated hydrocarbons such as dichloromethane, t
  • reaction is preferably carried out without solvent to the product (XITJ), in the event that R 2 and R 3 are both not hydrogen, the reaction is preferably in a mixture of toluene and acetonitrile to the product (XJJ) performed.
  • the usual inorganic or organic acids are suitable as acids for process step (X) + (XI) ⁇ (XU) or (XIII). These preferably include hydrochloric acid, sulfuric acid or phosphoric acid, or carboxylic acids such as formic acid, acetic acid or trifluoroacetic acid, or sulfonic acids such as toluenesulfonic acid, methanesulfonic acid or trifluoromethanesulfonic acid.
  • the usual Lewis acids such as boron trifluoride, aluminum trichlo- are also suitable. rid or zinc chloride. The acid is used in an amount of 1 to 10 mob based on 1 mol of the compound of formula (X).
  • reaction is preferably with 1 to 2 moles of zinc chloride to the product (XITJ), and in the event that R 2 and R 3 are both not hydrogen, preferably with 2 to 5 moles of trifluoroacetic acid to the product ( XU).
  • the reaction generally takes place in a temperature range from 0 ° C to + 250 ° C. If R 3 is hydrogen, the reaction is preferably carried out in a temperature range from + 130 ° C to + 200 ° C to the product (XTJI), if R 2 and R 3 are both not hydrogen, the reaction is preferred in a temperature range from 0 ° C to + 50 ° C to the product (XU).
  • the reaction can be carried out under normal, elevated or reduced pressure (for example from 0.5 to 5 bar). Generally one works at normal pressure.
  • Reducing agents suitable for process step (XU) or (XUI) -> (U) are boron, aluminum or silicon hydrides, such as, for example, borane, diborane, sodium borohydride, sodium cyano- borohydride, lithium aluminum hydride or triethylsilane, optionally in the presence of an acid or Lewis acid such as, for example, acetic acid, trifluoroacetic acid, aluminum trichloride or boron trifluoride, or hydrogenation with hydrogen in the presence of a suitable catalyst such as, for example, palladium on activated carbon, platinum oxide or Raney nickel.
  • a suitable catalyst such as, for example, palladium on activated carbon, platinum oxide or Raney nickel.
  • a suitable catalyst such as, for example, palladium on activated carbon, platinum oxide or Raney nickel.
  • Suitable solvents for process step (XU) or (XUI) - (U) are, for example, ethers such as diethyl ether, dioxane, tetrahydrofuran, glycol dimethyl ether or diethylene glycol dimethyl ether, alcohols such as methanol, ethanob n-propanob iso-propanob n-butanol or tert-butanob or Hydrocarbons such as benzene, xylobium toluene, hexane, cyclohexane or petroleum fractions, or other solvents such as acetonitrile, acetic acid or water. It is also possible to use mixtures of the solvents mentioned.
  • ethers such as diethyl ether, dioxane, tetrahydrofuran, glycol dimethyl ether or diethylene glycol dimethyl ether
  • alcohols such as methanol, ethanob n-propanob iso
  • Preferred for the reduction of the compounds of formula (XUI) is the use of acetic acid, which serves as an acid additive to the reducing agent in large excess at the same time as a solvent.
  • acetic acid which serves as an acid additive to the reducing agent in large excess at the same time as a solvent.
  • a mixture of methanol and toluene / acetonitrile [from the reaction (X) - (XU), with the addition of 2 to 5 mol of trifluoroacetic acid] in a ratio of 1: 1 to 1:10 used.
  • the reaction generally takes place in a temperature range from -20 ° C. to + 100 ° C., preferably from -10 ° C. to + 50 ° C.
  • the reaction can be carried out at normal, elevated or reduced pressure (e.g. from 0.5 to 5 bar). Generally one works at normal pressure.
  • the compounds of formula (I) according to the invention have a surprising and valuable pharmacological spectrum of activity and can therefore be used as versatile medicaments.
  • they are suitable for the treatment of coronary heart disease, for the prevention of myocardial infarction and for the treatment of restenosis after coronary angioplasty or stenting.
  • the compounds of formula (I) according to the invention are preferably suitable for the treatment of arteriosclerosis and hypercholesterolemia, for increasing morbidly low HDL levels and for lowering increased triglyceride and LDL levels. They can also be used to treat obesity, diabetes, metabolic syndrome (glucose intolerance, hyperinsulinemia, dyslipidaemia and high blood pressure due to insulin resistance), liver fibrosis and cancer.
  • the new active substances can be used alone or, if necessary, in combination with other active substances, preferably from the group CETP inhibitors, antidiabetics, antioxidants, cytostatics, calcium antagonists, antihypertensive agents, thyroid hormones and / or thyroid mimetics, inhibitors of HMG-CoA reductase, inhibitors of HMG-CoA reductase expression, squalene synthesis inhibitors, ACAT inhibitors, blood circulation promoting agents, platelet aggregation inhibitors, anticoagulants, angiotensin U-receptor antagonists, cholesterol absorption inhibitors, MTP inhibitors, aldolate inhibitor, redibrase inhibitor , Anoretics, lipase inhibitors and PPAR- ⁇ and / or PPAR- ⁇ agonists can be administered.
  • active substances preferably from the group CETP inhibitors, antidiabetics, antioxidants, cytostatics, calcium antagonists, antihypertensive agents, thyroid hormones and / or thyroid mimetics
  • the effectiveness of the compounds according to the invention can e.g. Check in vitro using the transactivation assay described in the sample section.
  • the activity of the compounds according to the invention in vivo can be e.g. check by the examinations described in the example section.
  • all customary application forms are possible, i.e. i.e. oral, parenteral inhalation, nasal, sublinguab rectal, external as e.g. transdermab or local such as for implants or stents.
  • parenteral administration intravenous, intramuscular or subcutaneous administration, for example as a subcutaneous depot, should be mentioned in particular.
  • Oral or parenteral administration is preferred.
  • Oral application is very particularly preferred.
  • the active ingredients can be administered alone or in the form of preparations.
  • Preparations suitable for oral administration include tablets, capsules, pellets, dragees, pills, granules, solid and liquid aerosols, syrups, emulsions, suspensions and solutions.
  • the active ingredient must be present in such an amount that a therapeutic effect is achieved.
  • the active ingredient in a concentration of 0.1 to 100 wt .-%, in particular in particular 0.5 to 90% by weight, preferably 5 to 80% by weight, are present.
  • the concentration of the active ingredient should be 0.5 to 90% by weight, ie the active ingredient should be present in amounts which are sufficient to achieve the dosage range indicated.
  • the active ingredients can be converted into the customary preparations in a manner known per se. This is done using inert, non-toxic, pharmaceutically suitable carriers, auxiliaries, solvents, vehicles, emulsifiers and / or dispersants.
  • auxiliaries include: water, non-toxic organic solvents such as e.g. Paraffins, vegetable oils (e.g. sesame oil), alcohols (e.g. ethanob glycerol), glycols (e.g. polyethylene glycol), solid carriers such as natural or synthetic rock flour (e.g. talc or silicates), sugar (e.g. milk sugar), emulsifiers and dispersants (e.g. polyvinylpyrrolidone) and lubricants (e.g. magnesium sulfate).
  • non-toxic organic solvents such as e.g. Paraffins, vegetable oils (e.g. sesame oil), alcohols (e.g. ethanob glycerol), glycols (e.g. polyethylene glycol), solid carriers such as natural or synthetic rock flour (e.g. talc or silicates), sugar (e.g. milk sugar), emulsifiers and dispersants (e.g. polyvinylpyrrol
  • tablets can of course also contain additives such as sodium citrate together with additives such as starch, gelatin and the like.
  • additives such as sodium citrate together with additives such as starch, gelatin and the like.
  • Aqueous preparations for oral administration can also be mixed with flavor enhancers or colorants.
  • doses of 0.001 to 5 mg / kg, preferably 0.005 to 3 mg / kg of body weight are preferably administered per 24 hours.
  • a mixture of 45 ml of toluene / acetonitrile (49: 1) is flushed with argon for 5 minutes and then mixed with 3.00 g (13.4 mmol) of 4-bromophenylhydrazine. Then 3.71 ml (48.1 mmol) of trifluoroacetic acid are slowly added, taking care that the temperature does not exceed 35 ° C. The temperature is then kept at 35 ° C. and a solution of 1.05 g (14.6 mmol) of isobutyraldehyde in 4 ml of toluene / acetonitrile (49: 1) is slowly added dropwise within 2 h. The mixture is stirred for 4 h at 35 ° C.
  • step d) 44 mg (0.15 mmol) of the indoline derivative from Example 1, step d) are dissolved in 2 ml of anhydrous dichloromethane and 0.024 ml (0.3 mmol) of triethylamine and 1.8 mg (0.015 mmol) of DMAP are added.
  • a solution of 50 mg (0.165 mmol) of the sulfonyl chloride from stage c) in 2 ml of dichloromethane is added dropwise and the mixture is stirred at RT for 24 h. Water and ethyl acetate are added and the aqueous phase is extracted three more times with ethyl acetate.
  • the United organic phases are dried over sodium sulfate and the solvent is removed. The residue is purified by means of preparative HPLC.
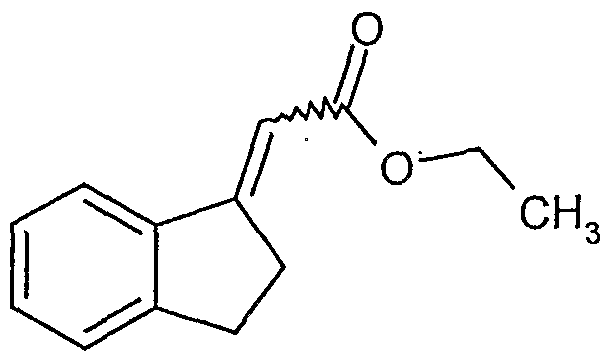
  • stage f) Analogously to the procedure for Example 1, stage f), 2.83 g (14 mmol) of the unsaturated ester from stage a) are hydrogenated on activated carbon in the presence of 10% palladium.
  • stage d 150 mg (0.515 mmol) of the indoline derivative from Example 1, stage d) are converted into 120 mg (40% of theory) by reaction with 311 mg (1.03 mmol) of the sulfonyl chloride from stage c) . Th.) Of the title compound.
  • a cellular assay is used to identify activators of the peroxisome proliferator-activated receptor delta (PPAR-delta).
  • the GAL4-PPAR ⁇ expression construct contains the ligand binding domain of PPAR ⁇ (amino acids 414-1326), which is PCR-amplified and cloned into the vector pcDNA3.1. This vector already contains the GAL4 DNA binding domain (amino acids 1-147) of the vector pFC2-dbd (Stratagene).
  • the reporter construct which contains five copies of the GAL4 binding site, upstream of a thymidine kinase promoter, leads to the expression of Firefly luciferase (Photinus pyralis) after activation and binding of GAL4-PPAR ⁇ .
  • CHO (Chinese hamster ovary) cells are in CHO-A-SFM medium (GTBCO) supplemented with 2.5% fetal calf serum and 1% penicillin / streptomycin (GEBCO), at a cell density 'of from 2 x 10 3 cells per well in a 384 well plate (Greiner) is sown. After culturing at 37 ° C. for 48 h, the cells are stimulated. For this purpose, the substances to be tested are taken up in the medium mentioned above and added to the cells. After a stimulation time of 24 hours, the luciferase activity is measured using a video camera. The measured relative light units result in a sigmoid stimulation curve depending on the substance concentration.
  • the EC S o values are calculated with the help of the computer program GraphPad PRISM (version 3.02). In this test, examples 1-3 show EC 50 values in a range from 10 to 100 nM.
  • the substances which are to be investigated for their HDL-C-increasing effect in vivo are administered orally to male transgenic hApoAl mice.
  • the substances are administered orally once a day for 7 days.
  • the test substances are dissolved in a solution of Solutol HS 15 + ethanol + saline (0.9%) in a ratio of 1 + 1 + 8 or in a solution of Solutol HS 15 + saline (0.9%) in a ratio of 2 + 8.
  • the dissolved substances are applied in a volume of 10 ml / kg body weight with a gavage. Animals that are treated in the same way but only receive the solvent (10 ml / kg body weight) without test substance serve as a control group.
  • blood is taken from each mouse for the determination of ApoAl, serum cholesterol, HDL-C and serum triglycerides (TG) by puncturing the retroorbital venous plexus (previous value).
  • the animals are then given the test substance for the first time using a pharyngeal tube.
  • 24 hours after the last substance application, i.e. on the 8th day after the start of treatment blood is again taken from each animal to determine the same parameters by puncturing the retroorbital venous plexus.
  • the blood samples are centrifuged and, after the serum has been obtained, cholesterol and TG are determined photometrically using an EPOS Analyzer 5060 (Eppendorf-Gedorfebau, Netheler & Hinz GmbH, Hamburg). The determination is carried out using commercially available enzyme tests (Boehringer Mannheim, Mannheim).
  • the non-HDL-C fraction is precipitated with 20% PEG 8000 in 0.2 M glycine buffer pH 10.
  • the cholesterol is determined from the supernatant in a 96-well perforated plate using a commercially available reagent (Ecoline 25, Merck, Darmstadt) using UV photometry (BIO-TEK Instruments, USA).
  • Human mouse ApoAl is sandwiched by ELISA using a polyclonal anti-human ApoAl and a monoclonal anti-human ApoAl antibody (Biode- sign Internationab USA).
  • the quantification is carried out by UV photometry (BIO-TEK Instruments, USA) with peroxidase-coupled anti-mouse IGG antibodies (KPL, USA) and peroxidase substrate (KPL, USA).
  • the effect of the test substances on the HDL-C concentration is determined by subtracting the measured value of the 1st blood sample (previous value) from the measured value of the 2nd blood sample (after treatment). The differences of all HDL-C values in a group are averaged and compared with the mean of the differences in the control group.
  • Substances that increase the HDL-C of the treated animals statistically significantly (p ⁇ 0.05) by at least 15% compared to that of the control group are considered to be pharmacologically active.
  • mice with insulin resistance and increased blood glucose levels are used.
  • C57B1 / 6J Lep ⁇ ob> mice are treated according to the same protocol as the transgenic ApoAl mice.
  • Serum lipids are determined as described above.
  • serum glucose is determined as a parameter for blood glucose in these animals. The serum glucose is determined enzymatically on an EPOS Analyzer 5060 (see above) using commercially available enzyme tests (Boehringer Mannheim).
  • a blood glucose-lowering effect of the test substances is determined by subtracting the measured value of the first blood sample from an animal (previous value) from the measured value of the second blood sample from the same animal (after treatment). The differences of all serum glucose values in a group are averaged and compared with the mean value of the differences in the control group.
  • the statistical evaluation is carried out with Student's t-test after the variances have been checked for homogeneity.
  • the compounds according to the invention can be converted into pharmaceutical preparations as follows:
  • the mixture of compound according to the invention, lactose and starch is granulated with a 5% solution (m / m) of the PVP in water. After drying, the granules are mixed with the magnesium stearate for 5 minutes. This mixture is compressed with a conventional tablet press (tablet format see above). A pressing force of 15 kN is used as a guideline for the pressing.
  • a single dose of 100 mg of the compound according to the invention corresponds to 10 ml of oral suspension. production:
  • Rhodigel is suspended in ethanol, the compound according to the invention is added to the suspension.
  • the water is added with stirring.
  • the mixture is stirred for about 6 hours until the swelling of the rhodel is complete.
  • the compound of the invention is suspended in the mixture of polyethylene glycol and polysorbate with stirring.
  • the stirring process is continued until the connection according to the invention has completely dissolved.

Landscapes

  • Health & Medical Sciences (AREA)
  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • Engineering & Computer Science (AREA)
  • Bioinformatics & Cheminformatics (AREA)
  • Veterinary Medicine (AREA)
  • Life Sciences & Earth Sciences (AREA)
  • Public Health (AREA)
  • Chemical Kinetics & Catalysis (AREA)
  • General Chemical & Material Sciences (AREA)
  • Medicinal Chemistry (AREA)
  • Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
  • General Health & Medical Sciences (AREA)
  • Pharmacology & Pharmacy (AREA)
  • Animal Behavior & Ethology (AREA)
  • Heart & Thoracic Surgery (AREA)
  • Cardiology (AREA)
  • Vascular Medicine (AREA)
  • Urology & Nephrology (AREA)
  • Child & Adolescent Psychology (AREA)
  • Diabetes (AREA)
  • Hematology (AREA)
  • Obesity (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Indole Compounds (AREA)

Abstract

Die vorliegende Anmeldung betrifft neue bicyclische Indolinsulfonamid-Derivate der allgemeinen Formel (I), Verfahren zu ihrer Herstellung sowie ihre Verwendung in Arzneimitteln, insbesondere als potente PPAR-delta aktivierende Verbindungen zur Prophylaxe und/oder Behandlung kardiovaskulärer Erkrankungen, insbesondere von Dyslipidämien, Arteriosklerose und koronaren Herzkrankheiten.

Description

Bicyclische lndolinsulfonamid-Derivate
Die vorliegende Anmeldung betrifft neue bicyclische Indolinsulfonamid-Derivate, Verfahren zu ihrer Herstellung sowie ihre Verwendung in Arzneimitteln, insbesondere als potente PPAR-delta aktivierende Verbindungen zur Prophylaxe und oder Behandlung kardiovaskulärer Erkrankungen, insbesondere von Dyslipidämien, Arteriosklerose und koronaren Herzkrankheiten.
Trotz vielfacher Therapieerfolge bleiben koronare Herzkrankheiten (KHK) ein ernstes Problem der öffentlichen Gesundheit. Während die Behandlung mit Statinen durch Hemmung der HMG-CoA- Reduktase sehr erfolgreich die Plasmakonzentration von LDL-Cholesterin senkt und dieses zu einer signifikanten Senkung der Mortalität von Risikopatienten fuhrt, so fehlen heute überzeugen- de Behandlungsstrategien zur Therapie von Patienten mit ungünstigem HDL/LDL-Cholesterin- Verhältnis und oder einer Hypertriglyceridämie.'
Fibrate stellen heute die einzige Therapieform für Patienten dieser Risikogruppen dar. Sie wirken als schwache Agonisten des Peroxisom-Proliferator-aktivierten Rezeptors (PPAR)-alpha (Nature 1990, 347, 645-50). Ein Nachteil von bisher zugelassenen Fibraten ist ihre nur schwache Interakti- on mit dem Rezeptor, die zu hohen Tagesdosen und deutlichen Nebenwirkungen führt.
Für den Peroxisom-Proliferator-aktivierten Rezeptor (PPAR)-delta (Mol. Endocrinok 1992, 6, 1634-41) weisen erste pharmakologische Befunde in Tiermodellen darauf hin, dass potente PPAR- delta-Agonisten ebenfalls zu einer Verbesserung des HDL/LDL-Cholesterin- Verhältnisses und der Hypertriglyceridämie führen können.
In der WO 00/23407 werden PPAR-Modulatoren zur Behandlung von Obesitas, Atherosklerose und/oder Diabetes offenbart. In der WO 93/15051 und EP 636 608-A1 werden 1-Benzolsulfonyl- l,3-dihydroindol-2-on-Derivate als Vasopressin- und/ oder Oxytocin-Antagonisten zur Behandlung verschiedener Erkrankungen beschrieben.
Aufgabe der vorliegenden Erfindung war die Bereitstellung neuer Verbindungen, die als PPAR- delta-Modulatoren eingesetzt werden können.
Gegenstand der vorliegenden Erfindung sind Verbindungen der allgemeinen Formel (I)
Figure imgf000002_0001
in welcher
R1 für Phenyl oder für 5- bis 6-gliedriges Heteroaryl mit bis zu zwei Heteroatomen aus der Reihe N, O und oder S steht, die ihrerseits jeweils ein- bis dreifach, gleich oder verschieden, durch Substituenten ausgewählt aus der Gruppe Halogen, Cyano, Nitro, (C C6)- Alkyl, welches seinerseits durch Hydroxy substituiert sein kann, (Cι-C6)-Alkoxy, Trifluor- methyl, Trifluormethoxy, (Cι-C6)-Alkylsulfonyb (C C6)-Alkanoyb (C C6)-Alkoxy- carbonyb Carboxyb Amino, (Cι-C6)-Acylamino, Mono- und Di-(Cι-C6)-alkylamino substituiert sein können,
R2 und R3 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder (Cj-C )- Alkyl stehen oder gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen 3- bis 7-gliedrigen, spiro-verknüpften Cycloalkyl-Ring bilden,
R4 für Wasserstoff oder (C, -C4)-Alkyl steht,
Rs und R6 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder (Cι-C4)- Alkyl stehen,
R7 für Wasserstoff oder für eine hydrolysierbare Gruppe steht, die zur entsprechenden Carbonsäure abgebaut werden kann,
und
n für die Zahl 1 oder 2 steht,
und ihre Salze, Solvate und Solvate der Salze.
Im Rahmen der Erfindung bedeutet in der Definition von R7 eine hydrolysierbare Gruppe eine Gruppe, die insbesondere im Körper zu einer Umwandlung der -C(0)OR7-Gruppierung in die entsprechende Carbonsäure (R7 = Wasserstoff) führt. Solche Gruppen sind beispielhaft und vorzugsweise: Benzyb (C C6)-Alkyl oder (C3-C8)-Cycloalkyb die jeweils gegebenenfalls ein- oder mehrfach, gleich oder verschieden, durch Halogen, Hydroxy, Amino, (C C6)-Alkoxy, Carboxyb (C C6)-Alkoxycarbonyb (Cι-C6)-Alkoxycarbonylamino oder (Cι-C6)-Alkanoyloxy substituiert sind, oder insbesondere (CrC4)-Alkyb das gegebenenfalls ein- oder mehrfach, gleich oder verschieden, durch Halogen, Hydroxy, Amino, (C C4)-Alkoxy, Carboxyb (C C4)-Alkoxycarbonyb (C C )- Alkoxycarbonylamino oder (C]-C )-Alkanoyloxy substituiert ist. Ci-CfiVAlkyl und (C C4)-Alkyl stehen im Rahmen der Erfindung für einen geradkettigen oder ver- zweigten Alkylrest mit 1 bis 6 bzw. 1 bis 4 Kphlenstoffatomen. Bevorzugt ist ein geradkettiger oder verzweigter Alkylrest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methyl, Ethyb n-Propyb Isopropyl und tert.-Butyb
(C Cg)-Cycloalkyl steht im Rahmen der Erfindung für eine monocyclische Cycloalkylgruppe mit 3 bis 8 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Cyclopropyb Cyclobutyb Cyclopentyl und Cyclohexyb
(Cr- )-Alkoxy und (Cv-CHAlkoxY stehen im Rahmen der Erfindung für einen geradkettigen oder verzweigten Alkoxyrest mit 1 bis 6 bzw. 1 bis 4 Kohlenstoffatomen. Bevorzugt ist ein geradkettiger oder verzweigter Alkoxyrest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methoxy, Ethoxy, n-Propoxy, Isopropoxy und tert.-Butoxy.
(C1-Cg)-Alkoxycarbonyl und ( -G -Alkoxycarbonyl stehen im Rahmen der Erfindung für einen geradkettigen oder verzweigten Alkoxyrest mit 1 bis 6 bzw. 1 bis 4 Kohlenstoffatomen, der über eine Carbonylgruppe verknüpft ist. Bevorzugt ist ein geradkettiger oder verzweigter Alkoxycarbonylrest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methoxycarbonyl, Ethoxycarbonyl, n-Propoxycarbonyb Isopropoxycarbonyl und tert-Butoxycarbonyb
( -CfiVAlkoxycarbonylamino und (C C^-Alkoxycarbonylamino stehen im Rahmen der Erfindung für eine Amino-Gruppe mit einem geradkettigen oder verzweigten Alkoxycarbonylsubsti- tuenten, der im Alkoxyrest 1 bis 6 bzw. 1 bis 4 Kohlenstoffatome aufweist und über die Carbonylgruppe verknüpft ist. Bevorzugt ist ein Alkoxycarbonylamino-Rest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methoxycarbonylamino, Ethoxycarbonylami- no, n-Propoxycarbonylamino und tert.-Butoxycarbonylamino.
fCq-Cg)-Alkanoyl steht im Rahmen der Erfindung für einen geradkettigen oder verzweigten Alkyl- Rest mit 1 bis 6 Kohlenstoffatomen, der in der 1 -Position ein doppelt gebundenes Sauerstoffatom trägt und über die 1 -Position verknüpft ist. Bevorzugt ist ein geradkettiger oder verzweigter Alka- noyl-Rest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Formyb Acetyb Propionyb n-Butyryb i-Butyryb Pivaloyl und n-Hexanoyb
(Cr-CsVAlkanoyloxy und (Cr-C bAlkanoyloxy stehen im Rahmen der Erfindung für einen geradkettigen oder verzweigten Alkyl-Rest mit 1 bis 6 bzw. 1 bis 4 Kohlenstoffatomen, der in der 1 -Position ein doppelt gebundenes Sauerstoffatom trägt und in der 1 -Position über ein weiteres Sauerstoffatom verknüpft ist. Bevorzugt ist ein Alkanoyloxy-Rest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Acetoxy, Propionoxy, n-Butyroxy, i-Butyroxy, Pivaloyloxy, n-Hexan- oyloxy. Mono-(C1-C≤ -Alkylamino und Mono-(C1-C4)-Alkylamino stehen im Rahmen der Erfindung für eine Amino-Gruppe mit einem geradkettigen oder verzweigten Alkylsubstituenten, der 1 bis 6 bzw. 1 bis 4 Kohlenstoffatome aufweist. Bevorzugt ist ein geradkettiger oder verzweigter Monoalkyl- amino-Rest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methyla- mino, Ethylamino, n-Propylamino, Isopropylamino und tert.-Butylamino.
Di-(C1-C≤)-All /lamino und DiJG-CaVAlkylamino stehen im Rahmen der Erfindung für eine Amino-Gruppe mit zwei gleichen oder verschiedenen geradkettigen oder verzweigten Alkylsubstituenten, die jeweils 1 bis 6 bzw. 1 bis 4 Kohlenstoffatome aufweisen. Bevorzugt sind geradkettige oder' verzweigte Dialkylamino-Reste mit jeweils 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugs- weise seien genannt: NN-Dimethylamino, N/V-Diethylamino, H-Ethyl-H-methylamino, N-Methyl-H- n-propylamino, TV-Isopropyl-N-n-propylamino, N-tert.-Butyl-H-memylamino, N-Ethyl-N-n-pentyl- amino und N-n-Hexyl-H-memylanjJ.no.
(C^-Cg -Acylamino steht im Rahmen der Erfindung für eine Amino-Gruppe mit einem geradkettigen oder verzweigten Alkanoyl-Substituenten, der 1 bis 6 Kohlenstoffatome aufweist und über die Car- bonylgruppe verknüpft ist. Bevorzugt ist ein Acylamino-Rest mit 1 bis 2 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Formamido, Acetamido, Propionamido, n-Butyramido und Pivaloylamido.
(C Cfi)-Alkylsulfonyl steht im Rahmen der Erfindung für einen geradkettigen oder verzweigten Alkylsulfonyl-Rest mit 1 bis 6 Kohlenstoffatomen. Bevorzugt ist ein geradkettiger oder verzweigter Alkylsulfonyl-Rest mit 1 bis 4 Kohlenstoffatomen. Beispielhaft und vorzugsweise seien genannt: Methylsulfonyb Ethylsulfonyb n-Propylsulfonyb Isopropylsulfonyb tert.-Butylsulfonyb n-Pentyl- sulfonyl und n-Hexylsulfonyb
5- bis 6-gliedriges Heteroaryl mit bis zu 2 gleichen oder verschiedenen Heteroatomen aus der Reihe N, O und/oder S steht im Rahmen der Erfindung für einen monocyclischen aromatischen Hetero- cyclus (Heteroaromaten), der über ein Ringkohlenstoffatom oder gegebenenfalls über ein Ringstickstoffatom des Heteroaromaten verknüpft ist. Beispielhaft seien genannt: Furanyb Pyrrolyb Thie- nyb Pyrazolyb Imidazolyb Thiazolyb Oxazolyb Isoxazolyb Isothiazolyb Pyridyb Pyrimidinyb Pyri- dazinyb Pyrazinyb Bevorzugt sind 5- bis 6-gliedrige Heteroaryl-Reste mit bis zu zwei Stickstoffatomen wie beispielsweise Imidazolyb Pyridyb Pyrimidinyb Pyridazinyb Pyrazinyb
Halogen schließt im Rahmen der Erfindung Fluor, Chlor, Brom und lod ein. Bevorzugt sind Chlor oder Fluor. Die erfindungsgemäßen Verbindungen können in Abhängigkeit von dem Substitutionsmuster in stereoisomeren Formen, die sich entweder wie Bild und Spiegelbild (Enantiomere), oder die sich nicht wie Bild und Spiegelbild (Diastereomere) verhalten, existieren. Die Erfindung betrifft sowohl die Enantiomeren oder Diastereomeren als auch deren jeweilige Mischungen. Die Racem- formen lassen sich ebenso wie die Diastereomeren in bekannter Weise in die stereoisomer einheitlichen Bestandteile trennen.
Weiterhin können bestimmte Verbindungen in tautomeren Formen vorliegen. Dies ist dem Fachmann bekannt, und derartige Verbindungen sind ebenfalls vom Umfang der Erfindung umfasst.
Die erfindungsgemäßen Verbindungen können auch als Salze vorliegen. Im Rahmen der Erfindung sind physiologisch unbedenkliche Salze bevorzugt.
Physiologisch unbedenkliche Salze können Salze der erfindungsgemäßen Verbindungen mit anorganischen oder organischen Säuren sein. Bevorzugt werden Salze mit anorganischen Säuren wie beispielsweise Chlorwasserstoffsäure, Bromwasserstoffsäure, Phosphorsäure oder Schwefelsäure, oder Salze mit organischen Carbon- oder Sulfonsäuren wie beispielsweise Essigsäure, Propionsäu- re, Maleinsäure, Fumarsäure, Äpfelsäure, Zitronensäure, Weinsäure, Milchsäure, Benzoesäure, oder Methansulfonsäure, Ethansulfonsäure, Benzolsulfonsäure, Toluolsulfonsäure oder Naphtha- lindisulfonsäure.
Physiologisch unbedenkliche Salze können ebenso Salze der erfindungsgemäßen Verbindungen mit Basen sein, wie beispielsweise Metall- oder Ammoniumsalze. Bevorzugte Beispiele sind Alka- limetallsalze (z.B. Natrium- oder Kaliumsalze), Erdalkalisalze (z.B. Magnesium- oder Calciumsal- ze), sowie Ammoniumsalze, die abgeleitet sind von Ammoniak oder organischen Aminen, wie beispielsweise Ethylamin, Di- bzw. Triethylamin, Ethyldiisopropylamin, Monoethanolamin, Di- bzw. Triethanolamin, Dicyclohexylamin, Dimethylaminoethanob Dibenzylamin, N-Methyl- morpholin, Dihydroabietylamin, 1-Ephenamin, Methylpiperidin, Arginin, Lysin, Ethylendiamin oder 2-Phenylethylamin.
Die erfindungsgemäßen Verbindungen können auch in Form ihrer Solvate, insbesondere in Form ihrer Hydrate vorliegen.
Bevorzugt sind Verbindungen der allgemeinen Formel (I), in welcher
R1 für Phenyl steht, das ein- bis zweifach, gleich oder verschieden, durch Substituenten aus- gewählt aus der Gruppe Fluor, Chlor, Cyano, (CrC4)-Alkyb (Cι-C4)-Alkoxy, Trifluor- methyb Trifluormethoxy, Methylsulfonyb '. Acetyb Propionyb (Cι-C4)-Alkoxycarbonyb Amino, Acetylamino, Mono- und Di-(Cι-C4)-alkylamino substituiert sein kann, R2 und R3 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder ( -C )- Alkyl stehen oder gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen 5- bis 6-gliedrigen, spiro-verknüpften Cycloalkyl-Ring bilden,
R4 für Wasserstoff oder Methyl steht,
R5 und R6 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder Methyl stehen,
R7 für Wasserstoff steht,
und
n für die Zahl 1 oder 2 steht.
Besonders bevorzugt sind Verbindungen der allgemeinen Formel (I), in welcher
R1 für Phenyl steht, das ein- bis zweifach, gleich oder verschieden, durch Substituenten ausgewählt aus der Gruppe Fluor, Chlor, Methyl, Trifluormethyl und Trifluormethoxy substituiert sein kann,
R2 für Methyl steht,
R3 für Methyl steht,
oder
R2 und R3 gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen spiro- verknüpften Cyclopentan- oder Cyclohexan-Ring bilden,
R4 für Wasserstoff oder Methyl steht,
R5 und R6 j eweils für Wasserstoff stehen,
R7 für Wasserstoff steht,
und
n für die Zahl 1 oder 2 steht.
Die oben aufgeführten allgemeinen oder in Vorzugsbereichen angegebenen Restedefinitionen gel- ten sowohl für die Endprodukte der Formel (I) als auch entsprechend für die jeweils zur Herstellung benötigten Ausgangsstoffe bzw. Zwischenprodukte. Die in den jeweiligen Kombinationen bzw. bevorzugten Kombinationen von Resten im einzelnen angegebenen Restedefinitionen werden unabhängig von den jeweilig angegebenen Kombinationen der Reste beliebig auch durch Restedefinitionen anderer Kombinationen ersetzt.
Ganz besonders bevorzugt sind Kombinationen von zwei oder mehreren der oben genannten Vor- zugsbereiche.
Von besonderer Bedeutung sind Verbindungen der Formel (I-A)
Figure imgf000008_0001
in welcher
R für Phenyl steht, das durch Fluor, Chlor oder Trifluormethyl substituiert ist,
und
n für die Zahl 1 oder 2 steht.
Außerdem wurde ein Verfahren zur Herstellung der erfϊndungsgemäßen Verbindungen der allgemeinen Formel (I) bzw. (I-A) gefunden, dadurch gekennzeichnet, dass man Verbindungen der Formel (U)
Figure imgf000008_0002
in welcher R , R und R jeweils die oben angegebenen Bedeutungen haben und
Y für Chlor oder Brom steht,
zunächst nach literaturüblichen Methoden in Verbindungen der Formel (r-Q)
Figure imgf000009_0001
in welcher Y, R , R und R jeweils die oben angegebenen Bedeutungen haben und
PG für eine geeignete Amino-Schutzgruppe, vorzugsweise für 4-Nitrophenylsulfonyl steht,
überführt, diese dann in einer Kupplungsreaktion mit einer Verbindung der Formel (IV)
Figure imgf000009_0002
in welcher R1 die oben angegebene Bedeutung hat und
R8 für Wasserstoff oder Methyl steht oder beide Reste zusammen eine CH2CH2- oder C(CH3)2-C(CH3)2-Brücke bilden,
in einem inerten Lösungsmittel in Gegenwart eines geeigneten Palladium-Katalysators und einer Base zu Verbindungen der Formel (V)
Figure imgf000009_0003
in welcher PG, R , R , R und R jeweils die oben angegebenen Bedeutungen haben,
umsetzt [vgl. z.B. W. Hahnfeld, M. Jung, Pharmazie 1994, 49, 18-20; idem, Liebigs Ann. Chem. 1994, 59-64], anschließend nach literaturüblichen Methoden die Schutzgruppe PG zu Verbin- düngen der Formel (VI)
Figure imgf000009_0004
in welcher R > lb τ R>2 , r R>3 und R jeweils die oben angegebenen Bedeutungen haben,
wieder entfernt, dann mit einer Verbindung der Formel (VII)
Figure imgf000010_0001
in welcher R >s , R und n jeweils die oben angegebenen Bedeutungen haben und
T für Benzyl oder (C C6)-Alkyl steht,
in einem inerten Lösungsmittel in Gegenwart einer Base in Verbindungen der Formel (VIII)
Figure imgf000010_0002
in welcher n, T, R , R , R , R , R und R jeweils die oben angegebenen Bedeutungen haben,
überführt, die Verbindungen der Formel (V1TJ) dann mit Säuren oder Basen oder im Falle, dass T für Benzyl steht, auch hydrogenolytisch zu den entsprechenden Carbonsäuren der Formel (LX)
Figure imgf000010_0003
in welcher n, R , R , R , R , R und R jeweils die oben angegebenen Bedeutungen haben,
umsetzt, gegebenenfalls diese Carbonsäuren (LX) nach bekannten Methoden zur Veresterung weiter zu Verbindungen der Formel (I) modifiziert, und die resultierenden Verbindungen der Formel (LX) bzw. (I) gegebenenfalls mit den entsprechenden (i) Lösungsmitteln und/oder (ii) Basen oder Säuren zu ihren Solvaten, Salzen und/oder Solvaten der Salze umsetzt.
Inerte Lösungsmittel für den Verfahrensschritt (ITJ) + (IV) -» (V) sind beispielsweise Ether wie Diethylether, Dioxan, Tetrahydrofuran, Glykoldimethylether oder Diethylenglykoldimethylether, Alkohole wie Methanol, Ethanol, n-Propanol, iso-Propanob n-Butanol oder tert.-Butanob Kohlenwasserstoffe wie Benzol, Xylob Toluob Hexan, Cyclohexan oder Erdölfraktionen, oder andere Lösungsmittel wie Dimethylformamid, Acetonitril oder auch Wasser. Ebenso ist es möglich, Gemische der genannten Lösungsmittel einzusetzen. Bevorzugt sind Toluob Dimethylformamid oder Acetonitril.
Als Basen für den Verfahrensschritt (HI) + (IV) -» (V) eignen sich die üblichen anorganischen oder organischen Basen. Hierzu gehören bevorzugt Alkalihydroxide wie beispielsweise Lithium-, Natrium- oder Kaliumhydroxid, Alkali- oder Erdalkalicarbonate wie Natrium-, Kalium- oder Cal- ciumcarbonat, Alkaliphosphate wie Natrium- oder Kaliumphosphat, oder organische Amine wie Pyridin, Triethylamin, Ethyldiisopropylamin, N-Methylmorpholin oder N-Methylpiperidin. Besonders bevorzugt sind Natrium- oder Kaliumcarbonat oder Kaliumphosphat.
Die Base wird hierbei in einer Menge von 1 bis 5, bevorzugt von 2 bis 3 Mob bezogen auf 1 Mol der Verbindung der Formel (TU) eingesetzt.
Geeignete Palladium-Katalysatoren für den Verfahrensschritt (ITJ) + (TV) -» (V) sind bevorzugt Palladium(O)- oder Palladium(lTj-Verbindungen, die präformiert eingesetzt werden, wie beispielsweise [1,1 '-Bis(diphenylphosphino)ferrocenyl]palladium(π)chlorid, Bis(triphenylphosphin)- palladium(π)chlorid oder Tetrakis(triphenylphosphin)palladium(0), oder die in situ aus einer geeigneten Palladiumquelle wie beispielsweise Bis(dibenzylidenaceton)palladium(0) und einem geeigneten Phosphinliganden erzeugt werden können.
Die Realction erfolgt im allgemeinen in einem Temperaturbereich von 0°C bis +150°C, bevorzugt von +20°C bis +120°C. Die Umsetzung kann bei normalem, erhöhtem oder bei erniedrigtem Druck durchgeführt werden (z.B. von 0.5 bis 5 bar). Im allgemeinen arbeitet man bei Normaldruck.
Inerte Lösungsmittel für den Verfahrensschritt (VI) + (VU) — > (Vl-LT) sind beispielsweise Halogenkohlenwasserstoffe wie Dichlormethan, Trichlormethan, Tetrachlormethan, Trichlorethan, Tetra- chlorethan, 1,2-Dichlorethan oder Trichlorethylen, Ether wie Diethylether, Dioxan, Tetrahydrofuran, Glykoldimethylether oder Diethylenglykoldimethylether, Kohlenwasserstoffe wie Benzol, Xylob Toluob Hexan, Cyclohexan oder Erdölfraktionen, oder andere Lösungsmittel wie Nitro- methan, Ethylacetat, Aceton, Dimethylformamid, Dimethylsulfoxid, Acetonitril, N-Methyl- pyrrolidinon oder Pyridin. Ebenso ist es möglich, Gemische der genannten Lösungsmittel einzusetzen. Bevorzugt sind Dichlormethan oder Tetrahydrofuran.
Als Basen für den Verfahrensschritt (VI) + (VII) -» (VITJ) eignen sich die üblichen anorganischen oder organischen Basen. Hierzu gehören bevorzugt Alkalihydroxide wie beispielsweise Lithium-, Natrium- oder Kaliumhydroxid, Alkali- oder Erdalkalicarbonate wie Natrium-, Kalium- oder Cal- ciumcarbonat, Alkalihydride wie Natriumhydrid, oder organische Amine wie Pyridin, Triethyl- amin, Ethyldiisopropylamin, N-Methylmorpholin oder N-Methylpiperidin. Besonders bevorzugt sind Aminbasen wie Triethylamin, Pyridin oder Ethyldiisopropylamin, gegebenenfalls in Gegen- wart katalytischer Mengen (ca. 10 mol-%) von 4-NN-Dimethylaminopyridin oder 4-Pyrrolidino- pyridin.
Die Base wird hierbei in einer Menge von 1 bis 5, bevorzugt von 1 bis 2.5 Mob bezogen auf 1 Mol der Verbindung der Formel (VIT) eingesetzt.
Die Reaktion erfolgt im allgemeinen in einem Temperaturbereich von -20°C bis +100°C, bevor- zugt von 0°C bis +75 °C. Die Umsetzung kann bei normalem, erhöhtem oder bei erniedrigtem
Druck durchgeführt werden (z.B. von 0.5 bis 5 bar). Im allgemeinen arbeitet man bei Normaldruck.
Inerte Lösungsmittel für den Verfahrensschritt (VITI) -» (LX) sind beispielsweise Halogenkohlenwasserstoffe wie Dichlormethan, 1,2-Dichlorethan oder Trichlorethylen, Ether wie Diethylether, Dioxan, Tetrahydrofuran, Glykoldimethylether oder Diethylenglykoldimethylether, Alkohole wie Methanol, Ethanob n-Propanob iso-Propanob n-Butanol oder tert.-Butanob Kohlenwasserstoffe wie Benzol, Xylob Toluob Hexan, Cyclohexan oder Erdölfraktionen, oder andere Lösungsmittel wie Nitromethan, Aceton, Dimethylformamid, Dimethylsulfoxid, Acetonitrib N-Methylpyrroli- dinon oder auch Wasser. Ebenso ist es möglich, Gemische der genannten Lösungsmittel einzusetzen. Bevorzugt sind Alkohole wie Methanol oder Ethanol und deren Gemische mit Tetrahydrofu- ran.
Als Basen für den Verfahrensschritt (VITJ) - (LX) eignen sich die üblichen anorganischen Basen. Hierzu gehören bevorzugt Alkalihydroxide wie beispielsweise Lithium-, Natrium- oder Kaliumhydroxid, oder Alkali- oder Erdalkalicarbonate wie Natrium-, Kalium- oder Calciumcarbonat. Besonders bevorzugt sind Lithium- oder Natriumhydroxid.
Die Base wird hierbei in einer Menge von 1 bis 5, bevorzugt von 1 bis 3 Mob bezogen auf 1 Mol der Verbindung der Formel (VWL) eingesetzt. Als Säuren für den Verfahrensschritt (NIJJ) -» (LX) eignen sich die üblichen anorganischen Säuren wie beispielsweise Salzsäure oder Schwefelsäure, oder Sulfonsäuren wie Toluolsulfonsäure, Me- thansulfonsäure oder Trifluormethansulfonsäure, oder Carbonsäuren wie Trifluoressigsäure.
Die Reaktion erfolgt im allgemeinen in einem Temperaturbereich von -20°C bis +100°C, bevor- zugt von 0°C bis +30°C. Die Umsetzung kann bei normalem, erhöhtem oder bei erniedrigtem
Druck durchgeführt werden (z.B. von 0.5 bis 5 bar). Im allgemeinen arbeitet man bei Normaldruck.
Die Verbindungen der Formel (11) sind bekannt oder können in Analogie zu literaturbekannten Verfahren beispielsweise dadurch hergestellt werden, dass man Verbindungen der Formel (X)
Figure imgf000013_0001
in welcher Y die oben angegebene Bedeutung hat,
in Gegenwart einer Säure oder Lewis-Säure, gegebenenfalls in einem inerten Lösungsmittel, mit einer Verbindung der Formel (XI)
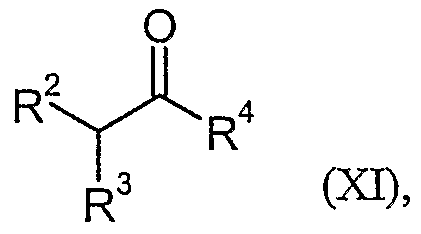
Figure imgf000013_0002
in welcher R , R und R jeweils die oben angegebenen Bedeutungen haben,
im Falb dass R2 und R3 in (XI) beide ungleich Wasserstoff sind, zu Verbindungen der Formel (XJI) bzw. im Falb dass R3 in (XI) für Wasserstoff steht, zu Verbindungen der Formel (XITJ)
Figure imgf000013_0003
(XU) (XIII)
in welchen Y und R4 jeweils die oben angegebenen Bedeutungen haben,
umsetzt und die Verbindungen der Formel (XII) bzw. (Xiπ) dann mit Hilfe eines Bor-, Alumini- um- oder Siliciumhydrids, wie beispielsweise Natriumborhydrid oder Natriumcyanoborhydrid, oder durch Hydrierung in Gegenwart eines geeigneten Katalysators, .wie beispielsweise Raney- Nickeb reduziert [für die Verfahrensschritte (X) + (XI) → (XU) → (IT) vgl. z.B. P.E. Maligres, I. Houpis, K. Rossen, A. Molina, b Sager, V. Upadhyay, K.M. Wells, R.A. Reamer, J.E. Lynch, D. Askin, R.P. Volante, P J. Reider, Tetrahedron 1997, 53, 10983-10992].
• Inerte Lösungsmittel für den Verfahrensschritt (X) + (XI) → (XU) bzw. (XITJ) sind beispielsweise Halogenkohlenwasserstoffe wie Dichlormethan, Trichlormethan, Tetrachlormethan, Trichlorethan, Tetrachlorethan, 1,2-Dichlorethan oder Trichlorethylen, Ether wie Dioxan, Tetrahydrofuran, Glykoldimethylether oder Diethylenglykoldimethylether, Alkohole wie Methanol, Ethanob n- Propanob iso-Propanob n-Butanol oder tert.-Butanob oder Kohlenwasserstoffe wie Benzol, Xylob Toluob Hexan, Cyclohexan oder Erdölfraktionen, oder andere Lösungsmittel wie Acetonitril oder Wasser. Ebenso ist es möglich, Gemische der genannten Lösungsmittel einzusetzen. Es ist auch möglich, die Reaktion ohne Lösungsmittel durchzuführen. Im Fall, dass R3 für Wasserstoff steht, wird die Reaktion bevorzugt ohne Lösungsmittel zum Produkt (XITJ) durchgeführt, im Fall, dass R2 und R3 beide ungleich Wasserstoff sind, wird die Reaktion bevorzugt in einem Gemisch aus Toluol und Acetonitril zum Produkt (XJJ) durchgeführt.
Als Säuren für den Verfahrensschritt (X) + (XI) → (XU) bzw. (XIII) eignen sich die üblichen anorganischen oder organischen Säuren. Hierzu gehören bevorzugt Salzsäure, Schwefelsäure oder Phosphorsäure, oder Carbonsäuren wie Ameisensäure, Essigsäure oder Trifluoressigsäure, oder Sulfonsäuren wie Toluolsulfonsäure, Methansulfonsäure oder Trifluormethansulfonsäure. Alterna- tiv eignen sich auch die üblichen Lewissäuren wie beispielsweise Bortrifluorid, Aluminiumtrichlo- . rid oder Zinkchlorid. Die Säure wird hierbei in einer Menge von 1 bis 10 Mob bezogen auf 1 Mol der Verbindung der Formel (X), eingesetzt. Im Falb dass R3 für Wasserstoff steht, wird die Reaktion bevorzugt mit 1 bis 2 Mol Zinkchlorid zum Produkt (XITJ), und im Fall, dass R2 und R3 beide ungleich Wasserstoff sind, bevorzugt mit 2 bis 5 Mol Trifluoressigsäure zum Produkt (XU) durch- geführt.
Die Reaktion erfolgt im allgemeinen in einem Temperaturbereich von 0°C bis +250°C. Im Falb dass R3 für Wasserstoff steht, wird die Reaktion bevorzugt in einem Temperaturbereich von +130°C bis +200°C zum Produkt (XTJI) durchgeführt, im Falb dass R2 und R3 beide ungleich Wasserstoff sind, wird die Reaktion bevorzugt in einem Temperaturbereich von 0°C bis +50°C zum Produkt (XU) durchgeführt. Die Umsetzung kann bei normalem, erhöhtem oder bei erniedrigtem Druck durchgeführt werden (z.B. von 0.5 bis 5 bar). Im allgemeinen arbeitet man bei Normaldruck.
Für den Verfahrensschritt (XU) bzw. (XUI) -> (U) geeignete Reduktionsmittel sind Bor-, Aluminium- oder Siliciumhydride, wie beispielsweise Boran, Diboran, Natriumborhydrid, Natriumcyano- borhydrid, Lithiumaluminiumhydrid oder Triethylsilan, gegebenenfalls in Gegenwart einer Säure oder Lewissäure wie beispielsweise Essigsäure, Trifluoressigsäure, Aluminiumtrichlorid oder Bortrifluorid, oder die Hydrierung mit Wasserstoff in Gegenwart eines geeigneten Katalysators wie beispielsweise Palladium auf Aktivkohle, Platinoxid oder Raney-Nickel. Bevorzugt ist bei Verbindungen der Formel (XUI) die Reduktion unter Verwendung von Natriumcyanoborhydrid; bei Verbindungen der Formel (XU) wird vorzugsweise Natriumborhydrid verwendet.
Geeignete Lösungsmittel für den Verfahrensschritt (XU) bzw. (XUI) — (U) sind beispielsweise Ether wie Diethylether, Dioxan, Tetrahydrofuran, Glykoldimethylether oder Diethylenglykoldimethylether, Alkohole wie Methanol, Ethanob n-Propanob iso-Propanob n-Butanol oder tert- Butanob oder Kohlenwasserstoffe wie Benzol, Xylob Toluob Hexan, Cyclohexan oder Erdölfraktionen, oder andere Lösungsmittel wie Acetonitril, Essigsäure oder Wasser. Ebenso ist es möglich, Gemische der genannten Lösungsmittel einzusetzen. Bevorzugt ist für die Reduktion der Verbindungen der Formel (XUI) die Verwendung von Essigsäure, die als Säurezusatz zum Reduktionsmittel im großen Überschuss gleichzeitig als Lösungsmittel dient. Für die Reduktion der Ver- bindungen der allgemeinen Formel (XU) wird vorzugsweise ein Gemisch aus Methanol und Tolu- ol/Acetonitril [aus der Umsetzung (X) - (XU), unter Zusatz von 2 bis 5 Mol Trifluoressigsäure] im Verhältnis 1 : 1 bis 1 : 10 verwendet.
Die Reaktion erfolgt im allgemeinen in einem Temperaturbereich von -20°C bis +100°C, bevorzugt von -10°C bis +50°C. Die Umsetzung kann bei normalem, erhöhtem oder bei erniedrigtem Druck durchgeführt werden (z.B. von 0.5 bis 5 bar). Im allgemeinen arbeitet man bei Normaldruck.
Die Verbindungen der Formel (VU) sind bekannt oder können in Analogie zu literaturbekannten Verfahren beispielsweise dadurch hergestellt werden, dass man Verbindungen der Formel (XIV)
Figure imgf000015_0001
in welcher n die oben angegebene Bedeutung hat,
zunächst in einer Wittig- oder Wittig-Horner-Reaktion [siehe z.B. J. Heterocycl. Chem. 1986, 747; J. Org. Chem. 1991, 6717] oder über Zinkorganyle [siehe z.B. J. Amer. Chem. Soc. 1958, 4360] in Verbindungen der Formel (XV)
Figure imgf000016_0001
in welcher R5, n und T jeweils die oben angegebenen Bedeutungen haben,
überführt, diese dann in Gegenwart eines geeigneten Katalysators, wie beispielsweise Palladium auf Aktivkohle, zu Verbindungen der Formel (XVI)
Figure imgf000016_0002
in welcher R5, n und T jeweils die oben angegebenen Bedeutungen haben,
hydriert und abschließend mit Chlorsulfonsäure umsetzt [vgl. z.B. P.D. Edwards, R.C. Mauger, K.M. CottreH, F.X. Morris, K.K. Pine, M.A. Sylvester, C.W. Scott, S.T. Furlong, Bioorg. Med. Chem. Lett. 2000, 10, 2291-2294].
Die Verbindungen der Formeln (IV), (X), (XI) und (XTV) sind kommerziell erhältlich, literaturbekannt oder können in Analogie zu literaturbekannten Verfahren hergestellt werden.
Das erfindungsgemäße Verfahren kann durch die folgenden Reaktionsschemata 1 und 2 veranschaulicht werden:
Schema 1
Figure imgf000017_0001
Figure imgf000017_0002
Schema 2
Figure imgf000018_0001
Figure imgf000018_0002
Die erfindungsgemäßen Verbindungen der Formel (I) zeigen ein überraschendes und wertvolles pharmakologisches Wiricirngsspektrum und lassen sich daher als vielseitige Medikamente einsetzen. Insbesondere eignen sie sich zur Behandlung der koronaren Herzkrankheit, zur Myokardin- farkt-Prophylaxe sowie zur Behandlung von Restenose nach Koronarangioplastie oder Stenting. Bevorzugt eignen sich die erfindungsgemäßen Verbindungen der Formel (I) zur Behandlung der Arteriosklerose und Hypercholesterolämie, zur Erhöhung krankhaft niedriger HDL-Spiegel sowie zur Senkung erhöhter Triglycerid- und LDL-Spiegel. Darüber hinaus können sie zur Behandlung von Obesitas, Diabetes, zur Behandlung des metabolischen Syndroms (Glucose-Intoleranz, Hype- rinsulinämie, Dyslipidämie und Bluthochdruck infolge von Insulinresistenz), der Leberfibrose und Krebs angewendet werden.
Die neuen Wirkstoffe können allein oder bei Bedarf in Kombination mit anderen Wirkstoffen vorzugsweise aus der Gruppe CETP-Inhibitoren, Antidiabetika, Antioxidantien, Cytostatika, Calcium- antagonisten, Blutdrucksenkende Mittel, Thyroidhormone und/oder Thyroidmimetika, Inhibitoren der HMG-CoA-Reduktase, Inhibitoren der HMG-CoA-Reduktase-Expression, Squalensynthese- Inhibitoren, ACAT-Inhibitoren, durchblutungsfördemde Mittel, Thrombozytenaggregations- hemmer, Antikoagulantien, Angiotensin-U-Rezeptorantagonisten, Cholesterin-Absorptions- hemmer, MTP-Inhibitoren, Aldolase-Redulttase-mhibitoren, Fibrate, Niacin, Anoretika, Lipase- Inhibitoren und PPAR-α- und/oder PPAR-γ-Agonisten verabreicht werden.
Die Wirksamkeit der erfindungs gemäßen Verbindungen lässt sich z.B. in vitro durch den im Bei- spielteil beschriebenen Transaktivierungsassay prüfen.
Die Wirksamkeit der erfindungsgemäßen Verbindungen in vivo lässt sich z.B. durch die im Beispielteil beschriebenen Untersuchungen prüfen.
Für die Applikation der Verbindungen der allgemeinen Formel (I) kommen alle üblichen Applikationsformen in Betracht, d.h. also oral, parenterab inhalativ, nasal, sublinguab rektal, äußerlich wie z.B. transdermab oder lokal wie z.B. bei Implantaten oder Stents. Bei der parenteralen Applikation sind insbesondere intravenöse, intramuskuläre oder subkutane Applikation, beispielsweise als subkutanes Depot, zu nennen. Bevorzugt ist die orale oder parenterale Applikation. Ganz besonders bevorzugt ist die orale Applikation.
Hierbei können die Wirkstoffe allein oder in Form von Zubereitungen verabreicht werden. Für die orale Applikation eignen sich als Zubereitungen u.a. Tabletten, Kapseln, Pellets, Dragees, Pillen, Granulate, feste und flüssige Aerosole, Sirupe, Emulsionen, Suspensionen und Lösungen. Hierbei muss der Wirkstoff in einer solchen Menge vorliegen, dass eine therapeutische Wirkung erzielt wird. Im allgemeinen kann der Wirkstoff in einer Konzentration von 0.1 bis 100 Gew.-%, insbe- sondere 0.5 bis 90 Gew.-%, vorzugsweise 5 bis 80 Gew.-%, vorliegen. Insbesondere sollte die Konzentration des Wirkstoffs 0.5 bis 90 Gew.-% betragen, d.h. der Wirkstoff sollte in Mengen vorliegen, die ausreichend sind, den angegebenen Dosierungsspielraum zu erreichen.
Zu diesem Zweck können die Wirkstoffe in an sich bekannter Weise in die üblichen Zubereitungen überführt werden. Dies geschieht unter Verwendung inerter, nichttoxischer, pharmazeutisch geeigneter Trägerstoffe, Hilfsstoffe, Lösungsmittel, Vehikel, Emulgatoren und/oder Dispergiermittel.
Als Hilfsstoffe seien beispielsweise aufgeführt: Wasser, nichttoxische organische Lösungsmittel wie z.B. Paraffine, pflanzliche Öle (z.B. Sesamöl), Alkohole (z.B. Ethanob Glycerin), Glykole (z.B. Polyethylenglykol), feste Trägerstoffe wie natürliche oder synthetische Gesteinsmehle (z.B. Talkum oder Silikate), Zucker (z.B. Milchzucker), Emulgiermitteb Dispergiermittel (z.B. Polyvi- nylpyrrolidon) und Gleitmittel (z.B. Magnesiumsulfat).
Im Falle der oralen Applikation können Tabletten selbstverständlich auch Zusätze wie Natriumcit- rat zusammen mit Zuschlagstoffen wie Stärke, Gelatine und dergleichen enthalten. Wässrige Zubereitungen für die orale Applikation können weiterhin mit Geschmacksaufbesserern oder Farbstof- fen versetzt werden.
Bei oraler Applikation werden vorzugsweise Dosierungen von 0.001 bis 5 mg/kg, bevorzugt von 0.005 bis 3 mg/kg Körpergewicht je 24 Stunden appliziert.
Die nachfolgenden Ausführungsbeispiele erläutern die Erfindung. Die Erfindung ist nicht auf die Beispiele beschränkt.
Die Prozentangaben in den folgenden Tests und Beispielen sind, sofern nicht anders angegeben, Gewichtsprozente; Teile sind Gewichtsteile. Lösungsmittelverhältnisse, Verdünnungsverhältnisse und Konzentrationsangaben von flüssig/flüssig-Lösungen beziehen sich jeweils auf das Volumen.
A. Beispiele
Verwendete Abkürzungen:
CI chemische Ionisation (bei MS)
DC Dünnschichtchromatographie
DCI direkte chemische Ionisation (bei MS)
DMAP 4-NN-Dimethylaminopyridin
DMF NN-Dimethylformamid
DMSO Dimethylsulfoxid d. Th. der Theorie (bei Ausbeute)
EI Elektronenstoß-Ionisation (bei MS)
ESI Elektrospray-Ionisation (bei MS)
Et Ethyb
GC Gaschromatographie h Stunde(n)
HPLC Hochdruck-, Hochleistungsflüssigchromatographie min Minute(n)
MS Massenspektroskopie
NMR Kernresonanzspektroskopie
Rf Retentionsindex (bei DC)
RT Raumtemperatur
THF Tetrahydrofuran
GC-MS-Methode:
Instrument: Micromass GCT, GC6890; Säule: Restek RTX-35MS, 30 x 250 μm x 0.25 μm; konstanter Fluss mit Helium: 0.88 ml/min; Ofen: 60°C; Inlet: 250°C; Gradient: 60°C (0.30 min halten), 50°C/min → 120°C, 16°C/min → 250°C, 30°C/min → 300°C (1.7 min halten). Ausfährungsbeispiele:
Beispiel 1
[7-({3,3-Dimethyb5-[4-(trifiuormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-b2,3,4- tetrahydro- 1 -naphthalinyl]essigsäure
Figure imgf000022_0001
Stufe a):
5 -Brom-3 ,3 -dimethylindolin
Figure imgf000022_0002
Ein Gemisch von 45 ml Toluol / Acetonitril (49:1) wird 5 Minuten mit Argon gespült und dann mit 3.00 g (13.4 mmol) 4-Bromphenylhydrazin versetzt. Anschließend fügt man langsam 3.71 ml (48.1 mmol) Trifluoressigsäure zu, wobei darauf geachtet wird, dass die Temperatur 35°C nicht überschreitet. Anschließend hält man die Temperatur auf 35°C und tropft langsam innerhalb von 2 h eine Lösung von 1.05 g (14.6 mmol) iso-Butyraldehyd in 4 ml Toluol / Acetonitril (49:1) zu. Man rührt 4 h bei 35°C und 2 h bei Raumtemperatur. Anschließend wird auf -10°C gekühlt, mit 4.0 ml Methanol versetzt und innerhalb von 30 min 819 mg (21.7 mmol) festes Natriumborhydrid in Portionen zugegeben. Dabei darf die Temperatur -2°C nicht überschreiten. Nach beendeter Zugabe wird 1 h bei 0°C gerührt. Nach Zugabe von 150 ml einer 6 Gew.-%-igen Lösung von Ammoniak in Wasser werden die Phasen getrennt und die organische Phase wird mit je 1.5 ml Acetonitril und Methanol versetzt. Anschließend wird die organische Phase mit 150 ml einer 15%-igen Lö- sung von Natriumchlorid in Wasser gewaschen und über Natriumsulfat getrocknet. Es wird durch 100 g Kieselgel filtriert und zweimal mit je 200 ml Diethylether nachgewaschen. Das organische Filtrat wird im Vakuum eingeengt und über 100 g Kieselgel chromatographiert. Zunächst werden mit Cyclohexan die Nebenprodukte eluiert, anschließend wird mit einem Gemisch Cyclohe- xan/Diethylether (20:1) das Produkt eluiert. Man erhält 1.78 g (54% d. Th.) des Produkts als Öl. Rf (Petrolether/Ethylacetat 5:1) = 0.47
MS (ESIpos): m/z = 226 [M+H]+
'H-NMR (200 MHz, DMSO-d6): δ = 1.20 (s, 6H), 3.18 (d, 2H), 5.66 (breites s, IH), 6.42 (d, IH), 7.02 (dd, IH), 7.10 (d, IH).
Stufe b):
5-Brom-3,3-dimethyl-l-(4-nitrophenylsulfonyl)-2,3-dihydro-7H-indol
Figure imgf000023_0001
17 g (75.18 mmol) des Bromindolins aus Stufe a), 5.22 g (150.37 mmol) Triethylamin und 0.46 g (3.76 mmol) DMAP werden in 100 ml Dichlormethan gelöst und auf 5-10°C abgekühlt. Man tropft eine Lösung von 17.5 g (78.94 mmol) 4-Nitrobenzolsulfonylchlorid in 150 ml Dichlormethan -zu und rührt bei RT für 16 h. Es wird jeweils einmal mit 2 M Salzsäure, Wasser und gesättigter Natri- umchlorid-Lösung gewaschen und die organische Phase über Natriumsulfat getrocknet. Nach Entfernen des Lösungsmittels erhält man 31 g (98 % d. Th.) des Produkts als gelben Feststoff.
MS (CI): m/z = 430 [M+NΗ4]+
Η-NMR (200 MHz, CDC13): δ = 8.32 (d, 2H), 8.0 (d, 2H), 7.51 (d, IH), 7.34 (d, IH), 7.15 (d, IH), 3.66 (s, 2H), 1.13 (s, 6H).
Stufe c):
3 , 3 -Dimethyl- 1 -(4-nitrophenylsulfonyl)-5 -[4-(trifluormethyl)phenyl] -2,3 -dihydroJH-indol
Figure imgf000023_0002
31 g (75.38 mmol) des geschützten Bromindolins aus Stufe b), 21.47 g (113.06 mmol) 4- (Trifluormethyl)phenylboronsäure und 15.63 g (113.06 mmol) Kaliumcarbonat werden in 500 ml Toluol suspendiert. Es wird für 30 min Argon durch die Lösung geleitet und danach 1.74 g (1.51 mmol) Tetrakis(triphenylphosphin)palladium(0) zugegeben. Man erhitzt für 16 h unter Rückfluss, kühlt dann ab und filtriert über eine ca. 1000 ml-Säule mit Kieselgel 60. Man eluiert erst mit ca. 1.5 1 Cyclohexan und danach mit ca. 2 1 Dichlormethan. Die Dichlormethan-Phase wird eingeengt. Man erhält 30 g (84 % d.Th.) des Produkts als gelben Feststoff.
MS (EI): m/z = 475.9 [M]+
'H-NMR (200 MHz, CDC13): δ = 8.32 (d, 2H), 8.05 (d, 2H), 7.71 (d, IH), 7.66 (d, 2H), 7.61 (d, 2H), 7.46 (dd, IH), 7.26 (s, IH), 3.73 (s, 2H), 1.21 (s, 6H).
Stufe d):
3,3-Dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-7H-indol
Figure imgf000025_0001
68 g (142.72 mmol) des Indolin-Derivats aus Stufe c) werden zusammen mit 25.12 g (0.628 mol) Natriumhydroxid in 300 ml DMF bei RT vorgelegt. 28.92 g (0.314 mol) Thioessigsäure werden zügig zugetropft und die Reaktionsmischung für 5 h auf 45°C erwärmt. Man gibt 1000 ml Essig- säureethylester zu und wäscht zweimal mit Sodalösung und einmal mit gesättigter Kochsalz- Lösung. Die organische Phase wird über Natriumsulfat getrocknet und eingeengt. Der Rückstand wird über Kieselgel 60 (1 kg) mit Cyclohexan/Essigsäureethylester (7:1) als Laufmittel filtriert. Man erhält 27.1 g (61 % d. Th.) des Produkts als hellgelb gefärbten Feststoff.
MS (EI): m z = 292.1 [M]+
Η-NMR (200 MΗz, CDCb): δ = 7.62 (s, 4Η), 7.31 (d, IH), 7.27 (m, 2H), 6.69 (d, IH), 3.39 (s, 2H), 1.36 (s, 6H).
Stufe e):
Ethyl 3,4-dihydro-l(2H)-naphthalinyTiden-ethanoat
Figure imgf000025_0002
In einem 100 ml-Rundkolben werden 13 g (40.22 mmol) Natriumethanolat vorgelegt und 6.7 ml (33.5 mmol) Triethylphosphonoacetat zugetropft. Es werden 10 ml wasserfreies Ethanol zugegeben und nach 5 min Rühren langsam 5 g (33.5 miftol) 1-Tetralon innerhalb von 10 min zugetropft. Man erhitzt die dunkelbraune Lösung unter Rückfluss für 18 h. Das Lösungsmittel wird vollständig entfernt und der Rückstand über Flashchromatographie an Kieselgel gereinigt (Laufmittel: Cyclohexan Essigsäureethylester 30+1 -» 7+1). Das so erhaltene Produkt (Gemisch der E/Z- Isomeren), das noch leicht mit 1-Tetralon verunreinigt ist, wird ohne .weitere Reinigung auf die unten beschriebene Weise hydriert.
Stufe f):
Ethyl 1,2,3 ,4-tetrahydro-l-naphthalinylacetat
Figure imgf000026_0001
4.2 g (19.4 mmol) des ungesättigten Esters aus Stufe e) werden in 300 ml Ethanol gelöst, 2 g Palladium auf Aktivkohle (10%-ig) zugegeben und für 4 h bei 3 bar Wasserstoffdruck hydriert. Der Katalysator wird abfiltriert und das Lösungsmittel entfernt. Der Rückstand wird im Ölpumpenva- kuum fraktioniert destilliert und die Fraktionen mittels GC-MS analysiert. Die Fraktion, die bei 160-170°C und ca. 1 mbar übergeht, enthält zu 99 % das gewünschte Produkt.
Ausbeute: 0.7 g (17 % d. Th.)
GC-MS (CI): m z = 219 [M+H]+
'H-NMR (200 MHz, DMSO-d6): δ = 7.18-6.99 (m, 4H), 4.09 (q, 2H), 3.20 (m, IH), 2.75-2.65 (m, 2H), 2.53-2.40 (m, 2H), 1.85-1.54 (m, 4H), 1.89 (t, 3H).
Stufe ε):
Ethyl [7-(chlorsulfonyl)-b2,3,4-tetrahydro-l-naphthalinyl]acetat
Figure imgf000026_0002
393 mg (1.787 mmol) des Tetrahydronaphthalin-Derivats aus Stufe f) werden in 13 ml Trichlor- methan bei 0°C vorgelegt und 831 mg (7.13 mmol) Chlorsulfonsäure zugetropft. Man rührt 3 h bei RT, gibt dann Eis zu und extrahiert dreimal mit Essigsäureethylester. Die vereinigten organischen Phasen werden über Magnesiumsulfat getrocknet und das Lösungsmittel vollständig entfernt. Das so erhaltene Rohprodukt wird ohne weitere Aufreinigung in der nächsten Stufe eingesetzt.
Ausbeute: 292 mg (52 % d. Th.)
MS (DCL NH3): m/z = 333 [M+NH3+NH4]+
'H-NMR (300 MHz, CDC13): δ = 7.69 (s, IH), 7.60 (d, IH), 7.12 (d, IH), 4.14 (q, 2H), 3.33-3.24 (m, IH), 2.89-2.48 (m, 4H), 1.90-1.65 (m, 4H), 1.24 (t, 3H).
Stufe h):
Ethyl [7-({3,3-dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-b2,3,4- tetrahydro- 1 -naphthalinyl]acetat
Figure imgf000027_0001
94 mg (0.324 mmol) des Indolin-Derivats aus Stufe d) werden in 3 ml Dichlormethan vorgelegt. Es werden 0.052 ml (0.65 mmol) Triethylamin und 4 mg (0.032 mmol) DMAP zugegeben. Bei RT tropft man eine Lösung von 113 mg (0.357 mmol) des Sulfonylchlorids aus Stufe g) in 2 ml Dichlormethan langsam zu und rührt 5 h bei RT. Das Lösungsmittel wird vollständig entfernt und der Rückstand durch präparative ΗPLC gereinigt. Das Produkt wird als gelber Feststoff isoliert.
Ausbeute: 129 mg (63 % d. Th.)
MS (DCI, NΗ3): m/z = 589 [M+NH4]+
'H-NMR (500 MHz, CDC13): δ = 7.70 (d, IH), 7.66-7.61 (m, 4H), 7.56 (d, IH), 7.45 (d, IH), 7.23 (s, 2H), 7.15 (d, IH), 4.13 (m, 2H), 3.68 (q, 2H), 3.34 (m, IH), 2.82-2.72 (m, 2H), 2.54-2.41 (m, 2H), 1.93-1.62 (m, 4H), 1.24 (t, 3H), 1.23 (s, 3H), 1.17 (s, 3H).
Stufe i):
[7-({3,3-Dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-mdol-l-yl}sulfonyl)-b2,3,4- tetrahydro- 1 -naphthalinyl] essigsaure
Figure imgf000028_0001
100 mg (0.175 mmol) des Esterderivats aus Stufe h) werden in 1 ml THF und 1 ml Methanol gelöst und dann eine Lösung von 8.4 mg (0.35 mmol) Lithiurnhydroxid in 0.4 ml Wasser zugegeben. Man rührt bei 50°C für 1 h und stellt dann die abgekühlte Lösung mit 1 M Salzsäure auf pH 5 ein. Man extrahiert dreimal mit Essigsäureethylester, trocknet die vereinigten organischen Phasen über Natriumsulfat und entfernt das Lösungsmittel vollständig. Der Rückstand wird mittels präparativer HPLC gereinigt.
Ausbeute: 45 mg (38 % d. Th.)
MS (DCL NH3): m/z = 561 [M+NH4]+
Η-NMR (300 MHz, DMSO-d6): δ = 12.27 (breites s, IH), 7.83 (d, 2H), 7.75 (d, 2H), 7.72 (d, IH), 7.61-7.50 (m, 4H), 7.26 (d, IH), 3.71 (s, 2H), 3.22 (m, IH), 2.73 (dd, 2H), 2.57-2.34 (m, 2H), 1.88- 1.53 (m, 4H), 1.15 (s, 6H).
Beispiel 2
[6-({3,3-Dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3-dihydro- lH-inden-1-ybjessigsäure
Figure imgf000028_0002
Stufe a):
Ethyl 2,3-dihydro-lH-inden-l -yliden-ethanoat
Figure imgf000029_0001
In einem 100 ml-Rundkolben werden 19.9 g (88.9 mmol) Triethylphosphonoacetat in 30 ml THF vorgelegt und portionsweise 3.9 g (97.9 mmol) Natriumhydrid (60%-ig in Mineralöl) zugegeben. Dabei soll die Temperatur nicht über 30°C steigen. Nach erfolgter Zugabe wird für 10 min gerührt und dann 4 g (29J mmol) 1-Indanon zugegeben. Man erhitzt die Lösung unter Rückfluss für 18 h. Nach dem Abkühlen wird Wasser zugegeben und dreimal mit Essigsäureethylester extrahiert. Die vereinigten organischen Phasen werden einmal mit Natriumchlorid-Lösung gewaschen und dann über Natriumsulfat getrocknet. Nach Entfernen des Lösungsmittels wird das Rohprodukt an Kieselgel chromatographiert (Laufmittel: Cyclohexan/Essigsäureethylester 1+0 -> 10+1). Man erhält 5.53 g (70 % d. Th.) des Produktes (als Gemisch der E/Z-Isomeren), das laut GC-MS ca. 76 % rein ist.
MS (DCI, NH3): m/z = 220 [M+NH4]+.
Stufe b):
Ethyl 2 ,3 -dihydro- lH-inden- 1 -ylacetat
Figure imgf000029_0002
Analog der Vorschrift zu Beispiel 1, Stufe f) werden 1.88 g (9.34 mmol) des ungesättigten Esters aus Stufe a) in Gegenwart von 10%-igem Palladium auf Aktivkohle hydriert.
Ausbeute: 1.84 g (90 % d. Th.)
GC-MS (CI): m/z = 222 [M+NΗ4]+
'H-NMR (200 MHz, CDC13): δ = 7.35-7.09 (m, 4H), 4.18 (q, 2H), 3.59 (m, IH), 3.0-2J0 ( , 3H), 2.50-2.30 ( , 2H), 1.85-1.65 ( , IH), 1.28 (t, 3H). Stufe c):
Ethyl [6-(chlorsulfonyl)-2,3-dihydro-lH-inden-l-yl]acetat
Figure imgf000030_0001
1.4 g (12.24 mmol) Chlorsulfonsäure werden in 15 ml Chloroform vorgelegt und auf 70°C er- wärmt. Man tropft bei dieser Temperatur eine Lösung von 500 mg (2.45 mmol) des Indan-Derivats aus Stufe b) zu und rührt für 30 min. Nach dem Abkühlen wird Wasser zugegeben und dreimal mit Essigsäureethylester extrahiert. Die vereinigten organischen Phasen werden über Magnesiumsulfat getrocknet. Nach Entfernen des Lösungsmittels wird das Rohprodukt ohne weitere Aufreinigung zur Sulfonamidbildung eingesetzt.
Ausbeute: 437 mg (59 % d. Th.)
MS (DCI, NΗ3): m/z = 320 [M+NH4]+.
Stufe d):
Ethyl [6-({3)3-dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3- dihydro- lH-inden- 1 -yljacetat
Figure imgf000030_0002
In 2 ml wasserfreiem Dichlormethan werden bei RT 44 mg (0.15 mmol) des Indolin-Derivats aus Beispiel 1, Stufe d) gelöst und 0.024 ml (0.3 mmol) Triethylamin und 1.8 mg (0.015 mmol) DMAP zugegeben. Eine Lösung von 50 mg (0.165 mmol) des Sulfonylchlorids aus Stufe c) in 2 ml Dichlormethan wird zugetropft und 24 h bei RT gerührt. Es wird Wasser und Essigsäureethylester zu- gegeben und die wässrige Phase noch dreimal mit Essigsäureethylester extrahiert. Die vereinigten organischen Phasen werden über Natriumsulfat getrocknet und das Lösungsmittel entfernt. Der Rückstand wird mittels präparativer HPLC gereinigt.
Ausbeute: 17 mg (20 % d. Th.)
MS (ESIpos): m z = 558 [M+H]
Η-NMR (500 MHz, CDC13): δ = 7.73-7.59 (m, 7H), 7.44 (d, IH), 7.32-7.20 (m, 2H), 4.15 (q, 2H), 3.69 (s, 2H), 3.59 (m, IH), 3.02-2.82 (m, 2H), 2J0 (dd, IH), 2.45-2.33 (m, 2H), 1.76 (m, IH), 1.26 (t, 3H), 1.21 (s, 3H), 1.20 (s, 3H).
Stufe e):
[6-({3,3-Dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3-dihydro- lH-inden-1-yl] essigsaure
Figure imgf000031_0001
16 mg (0.029 mmol) des Esterderivats aus Stufe d) werden in 0.5 ml TΗF und 0.5 ml Methanol gelöst und dann eine Lösung von 2 mg (0.08 mmol) Lithiumhydroxid in 0.1 ml Wasser zugegeben. Man rührt bei 50°C für 1 h und stellt dann die abgekühlte Lösung mit 1 M Salzsäure auf pH 5 ein. Man extrahiert dreimal mit Essigsäureethylester, trocknet die vereinigten organischen Phasen über Natriumsulfat und entfernt das Lösungsmittel vollständig.
Ausbeute: 14 mg (91 % d. Th.)
MS (DCL NΗ3): m/z = 547 [M+NH4]+
'H-NMR (300 MHz, CDC13): δ = 7.74-7.57 (m, 7H), 7.45 (dd, IH), 7.33-7.21 (m, 2H), 3.70 (d, 2H), 3.62 (m, IH), 3.0-2.88 (m, 2H), 2.73 (dd, IH), 2.52-2.40 (m, 2H), 1.81 (m, IH), 1.22 (s, 3H), 1.18 (s, 3H). Beispiel 3
[5-({3,3-Dimethyl-5-[4-(1xifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3-dihydro- lH-inden-2-yl]essigsäure
Figure imgf000032_0001
Stufe a):
Ethyl b3-dihydro-2H-inden-2-ylidenacetat
Figure imgf000032_0002
In einem 100 ml-Rundkolben werden 31.2 g (136.2 mmol) Triethylphosphonoacetat in 225 ml TΗF vorgelegt und portionsweise 5.99 g (149.8 mmol) Natriumhydrid (60%-ig in Mineralöl) zu- gegeben. Dabei soll die Temperatur nicht über 30°C steigen. Nach erfolgter Zugabe wird für 10 min gerührt und dann 6 g (45.4 mmol) 2-Indanon zugegeben. Man erhitzt die Lösung unter Rückfluss für 18 h. Nach dem Abkühlen wird Wasser zugegeben und dreimal mit Essigsäureethylester extrahiert. Die vereinigten organischen Phasen werden einmal mit Natriumchlorid-Lösung gewaschen und dann über Natriumsulfat getrocknet. Nach Entfernen des Lösungsmittels wird das Roh- produkt an Kieselgel chromatographiert (Laufmittel: Cyclohexan/Essigsäureethylester 10+1).
Ausbeute: 2.83 g (24 % d. Th.)
MS (EIpos): m/z = 202 [M]+
'Η-NMR (300 MΗz, CDC13): δ = 7.39 (d, 1Η), 7.30 (d, 1Η), 7.22 (m, 1Η), 7.14 (dt, 1Η), 6.69 (s, 1Η), 4.17 (q, 2Η), 3.51 (s, 2H), 3.45 (s, 2H), 1.28 (t, 3H). Stufe b):
Ethyl 2,3-dihydro-lH-inden-2-ylacetat
Figure imgf000033_0001
Analog der Vorschrift zu Beispiel 1, Stufe f) werden 2.83 g (14 mmol) des ungesättigten Esters aus Stufe a) in Gegenwart von 10%-igem Palladium auf Aktivkohle hydriert.
Ausbeute: 2.66 g (93 % d. Th.)
MS (EIpos): m/z = 204 [M]+
Η-NMR (300 MΗz, CDC13): δ = 7.22-7.09 (m, 4Η), 4.15 (q, 2H), 3.13 (dd, 2H), 2.89 (quin, IH), 2.65 (dd, 2H), 2.48 (d, 2H), 1.27 (t, 3H).
Stufe c):
Ethyl [5-(chlorsulfonyl)-2,3-dihydro-lH-inden-2-yl]acetat
Figure imgf000033_0002
2.65 g (12.97 mmol) des Indan-Derivats aus Stufe b) werden bei 0°C in Trichlormethan vorgelegt und 15.12 g (0.137 mol) Chlorsulfonsäure langsam zugetropft. Man rührt für 2 h bei RT und gießt das Reaktionsgemisch auf Eis. Man extrahiert dreimal mit Essigsäureethylester und trocknet die vereinigten organischen Phasen über Natriumsulfat. Nach Entfernen des Lösungsmittels wird das Rohprodukt ohne weitere Aufreinigung zur Sulfonamidbildung eingesetzt.
Ausbeute: 3.56 g (55 % d. Th.)
MS (DCb NΗ3): m/z = 320 [M+NH4]+
Η-NMR (300 MHz, CDC13): δ = 7.22-7.09 (m, 3H), 4.15 (q, 2H), 3.13 (dd, 2H), 2.89 (quin, IH), 2.65 (dd, 2H), 2.48 (d, 2H), 1.27 (t, 3H). Stufe d):
Ethyl [5-({3,3-dimethyl-5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3- dihydro-lH-inden-2-yl]acetat
Figure imgf000034_0001
Analog der Vorschrift zu Beispiel 2, Stufe d) werden aus 150 mg (0.515 mmol) des Indolin- Derivats aus Beispiel 1, Stufe d) durch Umsetzung mit 311 mg (1.03 mmol) des Sulfonylchlorids aus Stufe c) 120 mg (40 % d. Th.) der Titelverbindung hergestellt.
MS (DCI, NΗ3): m/z = 575 [M+NH4
Η-NMR (300 MHz, CDC13): δ = 7.72-7.58 (m, 7H), 7.43 (dd, IH), 7.28-7.22 (m, 2H), 4.12 (q, 2H), 3.70 (s, 2H), 3.15 (dd, 2H), 2.90 (quin, IH), 2.66 (dd, 2H), 2.45 (d, 2H), 1.26 (s, 3H), 1.25 (s, 3H), 1.23 (t, 3H).
Stufe e):
[5-({3,3-Dimethyb5-[4-(trifluormethyl)phenyl]-2,3-dihydro-lH-indobl-yl}sulfonyl)-2,3-dihydro- lH-inden-2-yl]essigsäure
Figure imgf000034_0002
100 mg (0.179 mmol) des Esterderivats aus Stufe d) werden in 5 ml TΗF und 5 ml Methanol gelöst und dann eine Lösung von 9 mg (0.359 mmol) Lithiumhydroxid in 1 ml Wasser zugegeben. Man rührt bei 50°C für 16 h und stellt dann die abgekühlte Lösung mit 2 M Salzsäure auf pH 5 ein. Man entfernt das Lösungsmittel vollständig und reinigt den Rückstand über präparative ΗPLC. Ausbeute: 90 mg (95 % d. Th.)
'H-NMR (300 MHz, DMSO-d6): δ = 12.06 (breites s, IH), 7.85 (d, 2H), 7J6 (d. 2H), 7J2 (s, IH), 7.66-7.52 (m, 4H), 7.39 (d, IH), 3.71 (s, 2H), 3.09 (m, 2H), 2.77-2.59 (m, 3H), 2.37 (d, 2H), 1.99 (s, 6H).
B. Bewertung der physiologischen Wirksamkeit
Beispiel A
Zellulärer Transaktivierungs-Assay:
Testprinzip:
Ein zellulärer Assay wird eingesetzt zur Identifizierung von Aktivatoren des Peroxisom- Proliferator-aktivierten Rezeptors delta (PPAR-delta).
Da Säugetierzellen verschiedene endogene nukleare Rezeptoren enthalten, die eine eindeutige Interpretation der Ergebnisse komplizieren könnten, wird ein etabliertes Chimärensystem eingesetzt, in dem die Liganden-Bindungsdomäne' des humanen PPARδ-Rezeptors an die DNA- Bindungsdomäne des Hefe-Transkriptionsfaktors GAL4 fusioniert wird. Die so entstehende GAL4-PPARδ-Chimäre wird in CHO- Zellen mit einem Reporterkonstrukt co-transfiziert und stabil exprimiert.
Klonierung:
Das GAL4-PPARδ-Expressions-Konstrukt enthält die Ligandenbindungsdomäne von PPARδ (A- minosäuren 414-1326), welche PCR-amplifiziert wird und in den Vektor pcDNA3.1 hineinkloniert wird. Dieser Vektor enthält bereits die GAL4-DNA-Bindungsdomäne (Aminosäuren 1-147) des Vektors pFC2-dbd (Stratagene). Das Reporterkonstrukt, welches fünf Kopien der GAL4- Bindestelle, vorgeschaltet vor einem Thymidinkinasepromoter enthält, führt zur Expression der Firefly-Luciferase (Photinus pyralis) nach Aktivierung und Bindung von GAL4-PPARδ.
Transaktivierungs-Assay (Luciferase-Reporter):
CHO (chinese hamster ovary)-Zellen werden in CHO-A-SFM-Medium (GTBCO), supplementiert mit 2.5% fötalem Kälberserum und 1% Penicillin/Streptomycin (GEBCO), mit einer Zelldichte ' von 2 x 10³ Zellen pro well in einer 384 well-Platte (Greiner) ausgesät. Nach Kultivierung über 48 h bei 37°C werden die Zellen stimuliert. Dazu werden die zu prüfenden Substanzen in oben ge- nanntem Medium aufgenommen und zu den Zellen dazu gegeben. Nach einer Stimulationszeit von 24 Stunden wird die Luciferaseaktivität mit Hilfe einer Videokamera gemessen. Die gemessenen relativen Lichteinheiten ergeben in Abhängigkeit von der Substanz-konzentration eine sigmoide Stimulationskurve. Die Berechnung der ECSo-Werte erfolgt mit Hilfe des Computerprogramms GraphPad PRISM (Version 3.02). Die Ausführungsbeispiele 1-3 zeigen in diesem Test EC50-Werte in einem Bereich von 10 bis 100 nM.
Beispiel B
Testbeschreibungen zur Auffindung von pharmakologisch wirksamen Substanzen, die das HDL- Cholesterin (HDL-C) im Serum von transgenen Mäusen, die' mit dem humanen ApoAl-Gen (hA- poAl) transfϊziert sind, erhöhen bzw. das metabolische Syndrom von adipösen ob,ob-Mäusen beeinflussen und deren Blutglucosekonzentration senken:
Die Substanzen, die auf ihre HDL-C erhöhende Wirkung in vivo untersucht werden sollen, werden männlichen transgenen hApoAl -Mäusen oral verabreicht. Die Tiere werden einen Tag vor Ver- suchsbeginn randomisiert Gruppen mit gleicher- Tierzahb in der Regel n = 7-10, zugeordnet. Während des gesamten Versuches steht den Tieren Trinkwasser und Futter ad libitum zur Verfügung. Die Substanzen werden einmal täglich 7 Tage lang oral verabreicht. Zu diesem Zweck werden die Testsubstanzen in einer Lösung aus Solutol HS 15 + Ethanol + Kochsalzlösung (0.9 %) im Verhältnis 1+1+8 oder in einer Lösung aus Solutol HS 15 + Kochsalzlösung (0.9 %) im Verhältnis 2+8 gelöst. Die Applikation der gelösten Substanzen erfolgt in einem Volumen von 10 ml/kg Körpergewicht mit einer Schlundsonde. Als Kontrollgruppe dienen Tiere, die genauso behandelt werden, aber nur das Lösungsmittel (10 ml/kg Körpergewicht) ohne Testsubstanz erhalten.
Vor der ersten Substanzapplikation wird jeder Maus zur Bestimmung von ApoAl, Serumcholeste- rin, HDL-C und Serumtriglyceriden (TG) Blut durch Punktion des retroorbitalen Venenplexus entnommen (Vorwert). Anschließend wird den Tieren mit einer Schlundsonde die Testsubstanz zum ersten Mal verabreicht. 24 Stunden nach der letzten Substanzapplikation, d.h. am 8. Tag nach Behandlungsbeginn, wird jedem Tier zur Bestimmung der gleichen Parameter erneut Blut durch Punktion des retroorbitalen Venenplexus entnommen. Die Blutproben werden zentrifugiert, und nach Gewinnung des Serums werden Cholesterin und TG photometrisch mit einem EPOS Analy- zer 5060 (Eppendorf-Gerätebau, Netheler & Hinz GmbH, Hamburg) bestimmt. Die Bestimmung erfolgt mit handelsüblichen Enzymtests (Boehringer Mannheim, Mannheim).
Zur Bestimmung des HDL-C wird die nicht-HDL-C-Fraktion mit 20% PEG 8000 in 0.2 M Glycin- puffer pH 10 gefällt. Aus dem Überstand wird das Cholesterin in einer 96er-Lochplatte mit handelsüblichem Reagenz (Ecoline 25, Merck, Darmstadt) UV-photometrisch bestimmt (BIO-TEK Instruments, USA).
Das humane Maus-ApoAl wird mit einer Sandwich-ELISA-Methode unter Verwendung eines polyklonalen antihuman- ApoAl- und eines monoklonalen antihuman- ApoAl -Antikörpers (Biode- sign Internationab USA) bestimmt. Die Quantifizierung erfolgt UV-phötometrisch (BIO-TEK Instruments, USA) mit Peroxidase-gekoppelten anti-Maus-IGG-Antikörper (KPL, USA) und Peroxi- dase-substrat (KPL, USA).
Die Wirkung der Testsubstanzen auf die HDL-C-Konzentration wird durch Subtraktion des Mess- wertes der 1. Blutentnahme (Vorwert) von dem Messwert der 2. Blutentnahme (nach Behandlung) bestimmt. Es werden die Differenzen aller HDL-C-Werte einer Gruppe gemittelt und mit dem Mittelwert der Differenzen der Kontrollgruppe verglichen.
Die statistische Auswertung erfolgt mit Student's t-Test nach vorheriger Überprüfung der Varianzen auf Homogenität.
Substanzen, die das HDL-C der behandelten Tiere, verglichen mit dem der Kontrollgruppe, statistisch signifikant (p<0.05) um mindestens 15% erhöhen, werden als pharmakologisch wirksam angesehen.
Um Substanzen auf ihre Beeinflussung eines metabolischen Syndroms prüfen zu können, werden Tiere mit einer Insulinresistenz und erhöhten Blutglucosespiegeln verwendet. Dazu werden C57B1/6J Lep <ob> - Mäuse nach dem gleichen Protokoll behandelt wie die transgenen ApoAl- Mäuse. Die Serumlipide werden wie oben beschrieben bestimmt. Zusätzlich wird bei diesen Tieren Serumglucose als Parameter für die Blutglucose bestimmt. Die Serumglucose wird enzymatisch an einem EPOS Analyzer 5060 (s. oben) mit handelsüblichen Enzymtests (Boehringer Mannheim) bestimmt.
Eine Blutglucose-senkende Wirkung der Testsubstanzen wird durch Subtraktion des Messwertes der 1. Blutentnahme eines Tieres (Vorwert) von dem Messwert der 2. Blutentnahme des gleichen Tieres (nach Behandlung) bestimmt. Es werden die Differenzen aller Serumglucose-Werte einer Gruppe gemittelt und mit dem Mittelwert der Differenzen der Kontrollgruppe verglichen.
Die statistische Auswertung erfolgt mit Student's t-Test nach vorheriger Überprüfung der Varian- zen auf Homogenität.
Substanzen, die die Serumglucosekonzentration der behandelten Tiere, verglichen mit der der Kontrollgruppe, statistisch signifikant (p<0.05) um mindestens 10% senken, werden als pharmakologisch wirksam angesehen. C. Ausführungsbeispiele für pharmazeutische Zusammensetzungen
Die erfindungsgemäßen Verbindungen können folgendermaßen in pharmazeutische Zubereitungen überführt werden:
Tablette:
Zusammensetzung:
100 mg der Verbindung von Beispiel 1, 50 mg Lactose (Monohydrat), 50 mg Maisstärke (nativ), 10 mg Polyvinylpyrrolidon (PVP 25) (Fa. BASF, Ludwigshafen, Deutschland) und 2 mg Magnesi- umstearat.
Tablettengewicht 212 mg. Durchmesser 8 mm, Wölbungsradius 12 mm.
Herstellung:
Die Mischung aus erfindungsgemäßer Verbindung, Lactose und Stärke wird mit einer 5%-igen Lösung (m/m) des PVPs in Wasser granuliert. Das Granulat wird nach dem Trocknen mit dem Magnesiumstearat 5 Minuten gemischt. Diese Mischung wird mit einer üblichen Tablettenpresse verpresst (Format der Tablette siehe oben). Als Richtwert für die Verpressung wird eine Presskraft von 15 kN verwendet.
Oral applizierbare Suspension:
Zusammensetzung:
1000 mg der Verbindung von Beispiel 1, 1000 mg Ethanol (96%), 400 mg Rhodigel® (Xanthan gum der Firma FMC, Pennsylvania, USA) und 99 g Wasser.
Einer Einzeldosis von 100 mg der erfindungsgemäßen Verbindung entsprechen 10 ml orale Suspension. Herstellung:
Das Rhodigel wird in Ethanol suspendiert, die erfindungsgemäße Verbindung wird der Suspension zugefügt. Unter Rühren erfolgt die Zugabe des Wassers. Bis zum Abschluß der Quellung des Rho- digels wird ca. 6 h gerührt.
Oral applizierbare Lösung:
Zusammensetzung:
500 mg der Verbindung von Beispiel 1, 2.5 g Polysorbat und 97 g Polyethylenglycol 400. Einer Einzeldosis von 100 mg der erfindungsgemäßen Verbindung entsprechen 20 g orale Lösung.
Herstellung:
Die erfindungsgemäße Verbindung wird in der Mischung aus Polyethylenglycol und Polysorbat unter Rühren suspendiert. Der Rührvorgang wird bis zur vollständigen Auflösung der erfmdungs- gemäßen Verbindung fortgesetzt.

Claims

Patentansprüche
1. Verbindungen der allgemeinen Formel (I)
Figure imgf000041_0001
in welcher
R1 für Phenyl oder für 5- bis 6-gliedriges Heteroaryl mit bis zu zwei Heteroatomen aus der Reihe N, O und/oder S steht, die ihrerseits jeweils ein- bis dreifach, gleich oder verschieden, durch Substituenten ausgewählt aus der Gruppe Halogen, Cya- no, Nitro,
Figure imgf000041_0002
welches seinerseits durch Hydroxy substituiert sein kann, (C C6)-Alkoxy, Trifluormethyl, Trifluormethoxy, (Cι-C6)-Alkylsulfonyl, (C C6)- Alkanoyb (C C6)-Alkoxycarbonyb Carboxyb Amino, (Cι-C6)-Acylamino, Mono- und Di-(C C6)-alkylamino substituiert sein können,
R2 und R3 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder (C]-C4)-Alkyl stehen oder gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen 3- bis 7-gliedrigen, spiro-verknüpften Cycloalkyl-Ring bilden,
R4 für Wasserstoff oder (C, -C4)-Alkyl steht,
R5 und R6 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder (Cι-C4)-Alkyl stehen,
R7 für Wasserstoff oder für eine hydrolysierbare Gruppe steht, die zur entsprechenden Carbonsäure abgebaut werden kann, und für die Zahl 1 oder 2 steht, sowie deren pharmazeutisch verträgliche Salze, Solvate und Solvate der Salze.
2. Verbindungen der allgemeinen Formel (I) gemäß Anspruch l, in welcher R1 für Phenyl steht, das ein- bis zweifach, gleich oder verschieden, durch Substituen- ten ausgewählt aus der Gruppe Fluor, Chlor, Cyano, (C C4)-Alkyb (CrC4)- Alkoxy, Trifluormethyb Trifluormethoxy, Methylsulfonyb Acetyb Propionyb ( - C )-Alkoxycarbonyb Amino, Acetylamino, Mono- und Di-(C C )-alkylamino sub- stituiert sein kann,
R2 und R3 gleich oder verschieden sind und unabhängig voneinander für Wasserstoff oder (Cj-O-Alkyl stehen oder gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen 5- bis 6-gliedrigen, spiro-verknüpften Cycloalkyl-Ring bilden,
R4 für Wasserstoff oder Methyl steht,
R5 und R6 gleich oder verschieden sind Und unabhängig voneinander für Wasserstoff oder Methyl stehen,
R7 für Wasserstoff steht,
und
n für die Zahl 1 oder 2 steht.
Verbindungen der allgemeinen Formel (I) gemäß Anspruch 1, in welcher
R1 für Phenyl steht, das ein- bis zweifach, gleich oder verschieden, durch Substituen- ten ausgewählt aus der Gruppe Fluor, Chlor, Methyl, Trifluormethyl und Trifluormethoxy substituiert sein kann,
R2 für Methyl steht,
R3 für Methyl steht,
oder
R2 und R3 gemeinsam mit dem Kohlenstoffatom, an das sie gebunden sind, einen spiro- verknüpften Cyclopentan- oder Cyclohexan-Ring bilden,
R4 für Wasserstoff oder Methyl steht,
R5 und R6 jeweils für Wasserstoff stehen,
R7 für Wasserstoff steht, und n für die Zahl 1 oder 2 steht.
4. Verbindungen der Formel (I-A)
Figure imgf000043_0001
in welcher
R für Phenyl steht, das durch Fluor, Chlor oder Trifluormethyl substituiert ist, und für die Zahl 1 oder 2 steht.
Verfahren zur Herstellung der Verbindungen der allgemeinen Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 4 definiert, dadurch gekennzeichnet, dass man Verbindungen der Formel (U)
Figure imgf000043_0002
in welcher R , R und R jeweils die in Anspruch 1 angegebenen Bedeutungen haben und
Y für Chlor oder Brom steht, zunächst nach literaturüblichen Methoden in Verbindungen der Formel (UJ)
Figure imgf000044_0001
in welcher Y, R2, R3 und R4 jeweils die in Anspruch 1 angegebenen Bedeutungen haben und
PG für eine geeignete Amino-Schutzgruppe, vorzugsweise für 4-Nitrophenylsulfonyl steht,
überführt, diese dann in einer Kupplungsreaktion mit einer Verbindung der Formel (TV)
Figure imgf000044_0002
in welcher R1 die in Anspruch 1 angegebene Bedeutung hat und
R8 für Wasserstoff oder Methyl steht oder beide Reste zusammen eine CH2CH2- oder C(CH3)2-C(CH3)2-Brücke bilden,
in einem inerten Lösungsmittel in Gegenwart eines geeigneten Palladium-Katalysators und einer Base zu Verbindungen der Formel (V)
Figure imgf000044_0003
in welcher PG, R1, R2, R3 und R4 jeweils die in Anspruch 1 angegebenen Bedeutungen haben,
umsetzt,- anschließend nach literaturüblichen Methoden die Schutzgruppe PG zu Verbindungen der Formel (VI)
Figure imgf000045_0001
in welcher R , R , R und R jeweils die in Anspruch 1 angegebenen Bedeutungen haben,
wieder entfernt, dann mit einer Verbindung der Formel (VU)
Figure imgf000045_0002
in welcher R , R und n jeweils die in Anspruch 1 angegebenen Bedeutungen haben und
T für Benzyl oder (C C6)-Alkyl steht,
in einem inerten Lösungsmittel in Gegenwart einer Base in Verbindungen der Formel
(vm)
Figure imgf000045_0003
in welcher n, T, R1, R2, R3, R4, R und Rδ jeweils die in Anspruch 1 angegebenen Bedeutungen haben,
überführt, die Verbindungen der Formel (VUI) dann mit Säuren oder Basen oder im Falle, dass T für Benzyl steht, auch hydrogenolytisch zu den entsprechenden Carbonsäuren der Formel (LX)
Figure imgf000046_0001
in welcher n, R1, R2, R3, R4, R und R jeweils die in Anspruch 1 angegebenen Bedeutungen haben, umsetzt, gegebenenfalls diese Carbonsäuren (LX) nach bekannten Methoden zur Ver- esterung weiter zu Verbindungen der Formel (I) modifiziert, und die resultierenden Verbindungen der Formel (LX) bzw. (I) gegebenenfalls mit den entsprechenden (i) Lösungsmitteln und/oder (ii) Basen oder Säuren zu ihren Solvaten, Salzen und/oder Solvaten der Salze umsetzt.
6. Verbindungen der Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 4 definiert, zur Vor- beugung und Behandlung von Krankheiten.
7. Arzneimittel enthaltend mindestens eine Verbindung der Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 4 definiert, und inerte, nichttoxische, pharmazeutisch geeignete Trägerstoffe, Hilfsmittel, Lösungsmittel, Vehikel, Emulgatoren und/oder Dispergiermittel.
8. Verwendung von Verbindungen der Formel (I) bzw. (I-A) und Arzneimittel, die in den Ansprüchen 1 bis 7 definiert sind, zur Vorbeugung vor und Behandlung von Krankheiten.
9. Verwendung von Verbindungen der Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 6 definiert, zur Herstellung von Arzneimitteln.
10. Verwendung von Verbindungen der Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 4 definiert, zur Herstellung von Arzneimitteln zur Vorbeugung und Behandlung von Stroke, Arteriosklerose, koronaren Herzkrankheiten und Dyslipidämie, zur Myokardinfarkt- Prophylaxe sowie zur Behandlung von Restenose nach Koronarangioplastie oder Stenting.
11. Verfahren zu Vorbeugung und Behandlung von Krankheiten,- dadurch gekennzeichnet, dass man Verbindungen der Formel (I) bzw. (I-A), wie in den Ansprüchen 1 bis 4 definiert, auf Lebewesen einwirken lässt.
PCT/EP2004/008075 2003-08-02 2004-07-20 Bicyclische indolinsulfonamid-derivate Ceased WO2005016881A1 (de)

Priority Applications (4)

Application Number Priority Date Filing Date Title
EP04763344A EP1654227A1 (de) 2003-08-02 2004-07-20 Bicyclische indolinsulfonamid-derivate
CA002534171A CA2534171A1 (en) 2003-08-02 2004-07-20 Bicyclic indoline sulfonamide derivatives
US10/566,340 US20070117860A1 (en) 2003-08-02 2004-07-20 Bicyclic indolinesulphonamide derivatives
JP2006522267A JP2007501195A (ja) 2003-08-02 2004-07-20 二環式インドリンスルホンアミド誘導体

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
DE10335449.2 2003-08-02
DE10335449A DE10335449A1 (de) 2003-08-02 2003-08-02 Bicyclische Indolinsulfonamid-Derivate

Publications (1)

Publication Number Publication Date
WO2005016881A1 true WO2005016881A1 (de) 2005-02-24

Family

ID=34072041

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/EP2004/008075 Ceased WO2005016881A1 (de) 2003-08-02 2004-07-20 Bicyclische indolinsulfonamid-derivate

Country Status (6)

Country Link
US (1) US20070117860A1 (de)
EP (1) EP1654227A1 (de)
JP (1) JP2007501195A (de)
CA (1) CA2534171A1 (de)
DE (1) DE10335449A1 (de)
WO (1) WO2005016881A1 (de)

Cited By (7)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO2007047432A1 (en) * 2005-10-12 2007-04-26 Kalypsys, Inc. Sulfonamide derivatives as modulators of ppar
WO2007051095A1 (en) 2005-10-25 2007-05-03 Kalypsys, Inc. Salts of modulators of ppar and methods of treating metabolic disorders
WO2008043024A3 (en) * 2006-10-04 2008-09-18 Kalypsys Inc Oral pharmaceutical formulation comprising a sulfonyl bicyclic modulator of ppar for the treatment of disease
US7557122B2 (en) 2005-09-01 2009-07-07 Laboratoires Fournier S.A. Pyrrolopyridine compounds, method of making them and uses thereof
US7576094B2 (en) 2004-12-13 2009-08-18 Eli Lilly And Company Spiro derivatives as lipoxygenase inhibitors
US7915253B2 (en) 2004-10-29 2011-03-29 Kalypsys, Inc Sulfonyl-substituted bicyclic compounds as modulators of PPAR
US9463189B2 (en) 2007-01-23 2016-10-11 Bpv Holdings, Llc Sulfonyl-substituted bicyclic compounds as PPAR modulators for the treatment of non-alcoholic steatohepatitis

Families Citing this family (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP1932843A1 (de) 2006-12-14 2008-06-18 sanofi-aventis Sulfonyl-Phenyl-2H-(1,2,4) Oxadiazol-5-one-Derivate, Verfahren zur Herstellung dafür und zur pharmazeutischen Verwendung damit
EP2288607B1 (de) 2008-06-09 2014-09-24 Sanofi Sulfonamide mit heterozyklen und einer oxadiazolon-kopfgruppe, verfahren zu ihrer herstellung und ihre verwendung als pharmazeutika
BRPI0914978A2 (pt) 2008-06-09 2015-10-27 Sanofi Aventis sulfonamidas n-heterocíclicas reforçadas com grupo principal de oxadiazolona, processos para sua preparação e seu uso como farmacêuticos

Citations (4)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP0636608A1 (de) * 1993-07-30 1995-02-01 Sanofi Derivate von 1-Benzolsulfonyl-1,3-dihydro-indol-2-on, ihre Herstellung und diese Verbindungen enthaltende pharmazeutische Zusammensetzungen
EP0769498A1 (de) * 1995-09-27 1997-04-23 Ono Pharmaceutical Co., Ltd. Sulfonamide mit Elastase inhibierender Wirkung
WO2002008188A1 (en) * 2000-07-25 2002-01-31 Merck & Co., Inc. N-substituted indoles useful in the treatment of diabetes
WO2003043985A1 (en) * 2001-11-21 2003-05-30 Novartis Ag Heterocyclic compounds and methods of use

Patent Citations (4)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP0636608A1 (de) * 1993-07-30 1995-02-01 Sanofi Derivate von 1-Benzolsulfonyl-1,3-dihydro-indol-2-on, ihre Herstellung und diese Verbindungen enthaltende pharmazeutische Zusammensetzungen
EP0769498A1 (de) * 1995-09-27 1997-04-23 Ono Pharmaceutical Co., Ltd. Sulfonamide mit Elastase inhibierender Wirkung
WO2002008188A1 (en) * 2000-07-25 2002-01-31 Merck & Co., Inc. N-substituted indoles useful in the treatment of diabetes
WO2003043985A1 (en) * 2001-11-21 2003-05-30 Novartis Ag Heterocyclic compounds and methods of use

Cited By (12)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US7915253B2 (en) 2004-10-29 2011-03-29 Kalypsys, Inc Sulfonyl-substituted bicyclic compounds as modulators of PPAR
US7576094B2 (en) 2004-12-13 2009-08-18 Eli Lilly And Company Spiro derivatives as lipoxygenase inhibitors
US7557122B2 (en) 2005-09-01 2009-07-07 Laboratoires Fournier S.A. Pyrrolopyridine compounds, method of making them and uses thereof
US7728002B2 (en) 2005-09-01 2010-06-01 Laboratoires Fournier S.A. Use of pyrrolopyridine compounds for activating PPAR receptors and treatment of conditions involving such receptors
WO2007047432A1 (en) * 2005-10-12 2007-04-26 Kalypsys, Inc. Sulfonamide derivatives as modulators of ppar
WO2007051095A1 (en) 2005-10-25 2007-05-03 Kalypsys, Inc. Salts of modulators of ppar and methods of treating metabolic disorders
JP2009513679A (ja) * 2005-10-25 2009-04-02 カリプシス・インコーポレーテッド Ppar調節剤の塩および代謝障害の治療方法
US7863276B2 (en) 2005-10-25 2011-01-04 Kalypsys, Inc Salts of modulators of PPAR and methods of treating metabolic disorders
CN101331122B (zh) * 2005-10-25 2011-09-07 凯利普西斯公司 Ppar调节剂的盐和治疗代谢性疾病的方法
KR101464767B1 (ko) * 2005-10-25 2014-11-24 칼립시스, 인코포레이티드 Ppar의 조절자의 염 및 대사질환을 치료하는 방법
WO2008043024A3 (en) * 2006-10-04 2008-09-18 Kalypsys Inc Oral pharmaceutical formulation comprising a sulfonyl bicyclic modulator of ppar for the treatment of disease
US9463189B2 (en) 2007-01-23 2016-10-11 Bpv Holdings, Llc Sulfonyl-substituted bicyclic compounds as PPAR modulators for the treatment of non-alcoholic steatohepatitis

Also Published As

Publication number Publication date
US20070117860A1 (en) 2007-05-24
JP2007501195A (ja) 2007-01-25
EP1654227A1 (de) 2006-05-10
DE10335449A1 (de) 2005-02-17
CA2534171A1 (en) 2005-02-24

Similar Documents

Publication Publication Date Title
DE10229777A1 (de) Indolin-Phenylsulfonamid-Derivate
EP1133486B1 (de) Nichtsteroidale entzündungshemmer
US11560369B2 (en) Crystalline forms and processes for the preparation of cannabinoid receptor modulators
DE69118388T2 (de) Substituierte bizyklische arylderivate mit selektiver leukotrien b4 antagonistischer wirkung
DE60309057T2 (de) Azabicycloderivate als antagonisten des muscarinischen rezeptors
DE10038639A1 (de) Nichtsteroidale Entzündungshemmer
EP1572671A1 (de) Nichtsteroidale entzündungshemmer
EP1583743A1 (de) Indol-phenylsulfonamid-derivate als ppar-delta aktivierende verbindungen
WO2002022586A1 (de) Indazole mit thyroid-hormon-ähnlicher wirkung, verfahren zu ihrer herstellung und ihre verwendung in arzneimitteln
WO2003097607A1 (de) Tetrahydroisochinolin-derivate
EP1654227A1 (de) Bicyclische indolinsulfonamid-derivate
DE10259244A1 (de) N-(Indolethyl-)cycloamin-Verbindungen
DD209446A5 (de) Verfahren zur herstellung von neuen aromatischen verbindungen
DE69317651T2 (de) Indol derivate als steroid 5 alpha-reductase inhibitoren
DE3850308T2 (de) Tetrahydroisochinolin-2-yl-Derivate als Thromboxan A2-Antagonisten.
DE69408674T2 (de) Phenylpyrrol-derivate und ihre verwendung als dopamin d3 antagonisten
EP1656134A1 (de) Indolin-sulfanilsäureamide als ppar-delta modulatoren
EP1675825A2 (de) Indolin-derivate als ppar delta modulatoren
DE19836406A1 (de) N-Substituierte Azabicycloheptan-Derivate, ihre Herstellung und Verwendung
DE102011104267A1 (de) Verbindungen als PPAR beta/delta Inhibitoren für die Behandlung von PPAR beta/delta-vermittelten Erkrankungen
EP2025673A1 (de) Arylsulfonamide mit analgetischer Wirkung
JPH072776A (ja) アンジオテンシンii拮抗性ピリジン誘導体
DE69711542T2 (de) Derivate von substituierten Benzoylharnstoffen und diese enthaltende Medikamente
DE102004049291A1 (de) Neue Oxadiazinon-Derivate und ihre Verwendung
DE10130830A1 (de) Indol-Derivate

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): AE AG AL AM AT AU AZ BA BB BG BR BW BY BZ CA CH CN CO CR CU CZ DE DK DM DZ EC EE EG ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX MZ NA NI NO NZ OM PG PH PL PT RO RU SC SD SE SG SK SL SY TJ TM TN TR TT TZ UA UG US UZ VC VN YU ZA ZM ZW

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): GM KE LS MW MZ NA SD SL SZ TZ UG ZM ZW AM AZ BY KG KZ MD RU TJ TM AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IT LU MC NL PL PT RO SE SI SK TR BF BJ CF CG CI CM GA GN GQ GW ML MR NE SN TD TG

121 Ep: the epo has been informed by wipo that ep was designated in this application
WWE Wipo information: entry into national phase

Ref document number: 2004763344

Country of ref document: EP

ENP Entry into the national phase

Ref document number: 2534171

Country of ref document: CA

WWE Wipo information: entry into national phase

Ref document number: 2006522267

Country of ref document: JP

WWP Wipo information: published in national office

Ref document number: 2004763344

Country of ref document: EP

WWE Wipo information: entry into national phase

Ref document number: 2007117860

Country of ref document: US

Ref document number: 10566340

Country of ref document: US

WWP Wipo information: published in national office

Ref document number: 10566340

Country of ref document: US