WO2008024692A1 - N-oxide aryl sulfones and sulfoxides - Google Patents
N-oxide aryl sulfones and sulfoxides Download PDFInfo
- Publication number
- WO2008024692A1 WO2008024692A1 PCT/US2007/076266 US2007076266W WO2008024692A1 WO 2008024692 A1 WO2008024692 A1 WO 2008024692A1 US 2007076266 W US2007076266 W US 2007076266W WO 2008024692 A1 WO2008024692 A1 WO 2008024692A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- compound
- alkyl
- hydrate
- salt
- independently chosen
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 CC(CC1)CC2(CC2)C*(C)N1C(COCC(C*(C)*CC(C)(C)C*1)N1S([Al])(=O)=O)=O Chemical compound CC(CC1)CC2(CC2)C*(C)N1C(COCC(C*(C)*CC(C)(C)C*1)N1S([Al])(=O)=O)=O 0.000 description 2
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D471/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00
- C07D471/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00 in which the condensed system contains two hetero rings
- C07D471/10—Spiro-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/92—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with a hetero atom directly attached to the ring nitrogen atom
- C07D211/94—Oxygen atom, e.g. piperidine N-oxide
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/92—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with a hetero atom directly attached to the ring nitrogen atom
- C07D211/96—Sulfur atom
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D213/00—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members
- C07D213/02—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members
- C07D213/89—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members with hetero atoms directly attached to the ring nitrogen atom
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D221/00—Heterocyclic compounds containing six-membered rings having one nitrogen atom as the only ring hetero atom, not provided for by groups C07D211/00 - C07D219/00
- C07D221/02—Heterocyclic compounds containing six-membered rings having one nitrogen atom as the only ring hetero atom, not provided for by groups C07D211/00 - C07D219/00 condensed with carbocyclic rings or ring systems
- C07D221/20—Spiro-condensed ring systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D413/00—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D413/02—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings
- C07D413/12—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings linked by a chain containing hetero atoms as chain links
Definitions
- This invention relates generally to N-oxide aryl sulfones and sulfoxides, and to the use of such compounds to treat conditions responsive to bradyki ⁇ iii receptor-1 (B ⁇ ) modulation.
- the invention further relates to the use of such compounds as reagents for the identification of other agents that bind to B 5 , and as probes for the detection and localization of B].
- Bradykinin is a nonapeptide that functions in cardiovascular homeostasis, contraction and relaxation of smooth muscles, inflammation and pain.
- the effects of BK are mediated by specific G protein-coupled BK receptors, of which there are at least two distinct subtypes termed Bj and B 2 .
- Bj and B 2 The B 2 receptor is expressed constitutively in a variety of tissues.
- the Bj receptor is inducibly expressed in response to pathophysiological conditions such as inflammation, pain, trauma, bacteria! infection, burns and shock.
- Bi is a particularly attractive drug target for these and other conditions, and agents that act at this receptor may be targeted specifically to injured tissues, with minimal effects in normal tissues.
- compounds that bind to Bi and/or modulate the activity of Bi also find use as research tools.
- the present invention provides N-oxide aryl sulfones and sulfoxides that satisfy Formula I:
- Formuia I or are a pharmaceutically acceptable salt, solvate (e.g., hydrate) or ester of such a compound.
- Ar is phenyl or a 5- or 6-membered heteroaryl. each of which is optionally substituted, and each of which is preferably substituted with from 0 to 4 substituents independently chosen from R s ; R x and R ⁇ are: fi) independently chosen from: (a) hydrogen; and
- W is CrQalkylene that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R 2 o;
- Z is absent, divalent phenyl, NR G or CR E R F wherein:
- R G is hydrogen, Cj-C 6 alkyl or taken together with R B or R D to form a 4- to 7-membered heterocycloalkyl that includes W and is optionally substituted; preferably such heterocycloalkyl is substituted with from 0 to 4 substituents independently chosen from R 20 ;
- R E is hydrogen, Ci-Qalkyl or taken together with R 8 to form a 4- to 7-membered heterocycloalkyl that includes W and is optionally substituted; preferably such heterocycloalkyl is substituted with from 0 to 4 substituents independently chosen from R 20 ; and
- R F is hydrogen, or taken together with R D to form a 4- to 7-membered heterocycloalkyl that includes V and is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R 20 ;
- V is absent or Cj-C 6 a!ky!ene that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from (i) R 20 and (ii) groups that are taken together with R 0 to form a 4- to 7-membered heterocycle that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R 2 o;
- is independently chosen from:
- R B is: (i) hydrogen, C,-C 6 alkyl, C 2 -C 6 alkenyl, C 2 -C 6 alkyny!, C r C 6 haloalkyl, (C 3 -C 3 cycloalky])Co-
- Rc is absent, C r C 6 alkyl or taken together with R D to from a 4- to 7-menibered heterocycle that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R 2 0;
- R D is CpQalkyl or taken together with Rc, R F , R G or a substituent of V to form an optionally substituted 4- to 7-membered heterocycle; and
- Each R 2O is independently selected from:
- Ciocarbocycle Co-C 4 alkyl and (4- to 10-membered heterocycle)C 0 -C 4 alkyl.
- N-oxide aryl sulfones and sulfoxides of Formula I, and other Formulas provided herein are B t modulators and exhibit a K 1 of no greater than 5 micromolar, 2 micrornolar, 1 micromolar, 500 nanomolar, 100 nanomolar, 50 nanomolar or 10 nanomolar in a Bi binding assay and/or have an EC S0 or IC 50 value of no greater than 5 micromolar, 2 micromolar, 1 micromolar, 500 nanomolar, 100 nanomolar, 50 nanomolar or 10 nanomolar in an assay for determination of B ⁇ agonist or antagonist activity.
- B 3 modulators provided herein are Bi antagonists; preferably such antagonists exhibit no detectable B] agonist activity,
- N-oxide aryl sulfones and sulfoxides of Formula I are labeled with a detectable marker (e.g., radiolabeled or fluorescein conjugated).
- the present invention further provides, within other aspects, pharmaceutical compositions comprising at least one N-oxide aryl sulfone or sulfoxide of Formula I in combination with a physiologically acceptable carrier or excipient.
- Methods are further provided for inhibiting agonist-induced B 1 activity.
- the inhibition takes place in vitro.
- Such methods comprise contacting a B
- the B 5 receptor is in a patient.
- Such methods comprise contacting cells expressing a B] receptor in a patient with at least one Bi antagonist as described herein in an amount or concentration that would be sufficient to detectably inhibit agonist-induced Bi activity in cells expressing a cloned B] receptor in vitro.
- the present invention further provides methods for treating a condition responsive to Bj receptor modulation in a patient, comprising administering to the patient a therapeutically effective amount of at least one N-oxide aiyl sulfone or sulfoxide of Formula I.
- methods for treating pain in a patient, comprising administering to a patient suffering from (or at risk for) pain a therapeutically effective amount of at least one N-oxide aryl sulfone or sulfoxide of Formula ⁇ .
- Pain conditions that may be treated include, but are not limited to, inflammatory pain, acute pain, dental pain, back pain, surgical pain, headache, neuropathic pain, and pain associated with osteoarthritis or trauma.
- the present invention provides methods for determining the presence or absence of Bi in a sample, comprising: (a) contacting a sample with an N-oxide aiyl sulfone or sulfoxide of Formula I under conditions that permit binding of the compound to B,; and (b) detecting a signal indicative of a level of the compound bound to Bi.
- the present invention provides methods of preparing the compounds disclosed herein, including the intermediates.
- the present invention provides N-oxide aryl suifones and sulfoxides, which may be used in vitro or in vivo in a variety of contexts, as described herein.
- N-oxide aryl sulfone or sulfoxide encompasses all compounds that satisfy Formula I, including compounds in which the Ar moiety is heteroaryl and compounds in which the Ar moiety is aiyl. This term further includes pharmaceutically acceptable salts, solvates (e.g., hydrates) and esters of such compounds.
- a "pharmaceutically acceptable salt” of a compound recited herein is an acid or base salt that is suitable for use in contact with the tissues of human beings or animals without excessive toxicity or carcinogenicity, and preferably without irritation, allergic response, or other problem or complication.
- Such salts include mineral and organic acid salts of basic residues such as amines, as well as alkali or organic salts of acidic residues such as carboxylic acids.
- Specific pharmaceutically acceptable anions for use in salt formation include, but are not limited to, acetate, 2-acetoxybenzoate, ascorbate, benzoate, bicarbonate, bitartrate, bromide, calcium edetate, carbonate, chloride, citrate, dihydrochloride, diphosphate, edetate, estolate (ethylsuccinate), formate, fumarate, gluceptate, gluconate, glutamate, glycolate, glycollylarsanilate, hexylresorcinate, hydrabamine, hydrobromide, hydrochloride, hydroiodide, hydroxymaleate, hydroxynaphthoate, iodide, isethionate, lactate, lactobionate, malate, maleate, mandelate, methyl bromide
- pharmaceutically acceptable cations for use in salt formation include, but are not limited to ammonium, benzathine, chloroprocaine, choline, diethanolamine, ethyl enediamine, meglumine, procaine, and metals such as aluminum, calcium, lithium, magnesium, potassium, sodium and zinc.
- metals such as aluminum, calcium, lithium, magnesium, potassium, sodium and zinc.
- a pharmaceutically acceptable acid or base salt can be synthesized from a parent compound that contains a basic or acidic moiety by any conventional chemical method.
- such salts can be prepared by reacting the free acid or base forms of these compounds with a stoichiometric amount of the appropriate base or acid in water or in an organic solvent, or in a mixture of the two; generally, the use of nonaqueous media, such as ether, ethyl acetate, ethanol, methanol, isopropanol or acetonitrile, is preferred.
- nonaqueous media such as ether, ethyl acetate, ethanol, methanol, isopropanol or acetonitrile
- each compound provided herein may, but need not, be formulated as a solvate (e.g., a hydrate) or non-covalent complex.
- the various crystal forms and polymorphs are within the scope of the present invention.
- prodrugs of the compounds provided herein are also provided herein.
- prodrug is a compound that may not fully satisfy the structural requirements of a formula provided herein, but is modified in vivo, following administration to a patient, to produce a compound within the scope of such formula.
- a prodrug may be an acylated derivative of a compound as provided herein.
- Prodrugs include compounds wherein hydroxy, amine or sulfhydryl groups are bonded to any group that, when administered to a mammalian subject, cleaves to form a free hydroxy, amino, or suifhydryl group, respectively.
- Examples of prodrugs include, but are not limited to, acetate, formate, phosphate and benzoate derivatives of alcohol and amine functional groups within the compounds provided herein.
- Prodrugs of the compounds provided herein may be prepared by modifying functional groups present in the compounds in such a way that the modifications are cleaved in vivo to yield the parent compounds.
- alkyl refers to a straight or branched chain saturated aliphatic hydrocarbon.
- Alkyl groups include groups having from 1 to 8 carbon atoms (C r C s alkyl), from 1 to 6 carbon atoms (C r C 6 aiky! and from 1 to 4 carbon atoms (Ci-C 4 alkyl) 3 such as methyl, ethyl, propyl, isopropyl, n-butyl, sec-butyl, / ⁇ ?r/-butyi, pentyl, 2-pentyl, isopentyl, neopentyl, hexyi. 2-hexyl, 3-hexyl or 3-metliylpentyl.
- C 0 -C 4 alkyr' refers to a single covalent bond (C 0 ) or an alkylene group having 1, 2, 3 or 4 carbon atoms;
- C 0 -C 2 alky! refers to a single covalent bond or a methylene or ethylene group.
- Alkenyl refers to straight or branched chain alkene groups, which comprise at least one unsaturated carbon-carbon double bond. Alkenyl groups include C 2 -Qalkenyl, C 2 -C 6 alkenyl and C 2 - C 4 alkenyl groups, which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively, such as ethenyl, ailyl or isopropenyl.
- AlkynyP refers to straight or branched chain alkyne groups, which have one or more unsaturated carbon-carbon bonds, at least one of which is a triple bond.
- Alkynyl groups include C 2 -Cgalkynyl, C 2 -C 6 aikynyl and C 2 -C 4 alkynyl groups, which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively.
- a “cycloaikyl” is a saturated or partially saturated cyclic group in which all ring members are carbon, such as cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl and partially saturated variants thereof. Certain cycloaikyl groups are C 3 -Cscycloalkyl, in which the ring contains from 3 to 8 ring members, all of which are carbon.
- a "(C 3 -C 3 cyc]oalkyI)C 0 -C 4 alkyr is a C 3 -Cgcycloalkyl group linked via a single covalent bond or a C r C 4 alkyIene group.
- alkoxy is meant an alkyl group attached via an oxygen bridge (i.e., -O- alkyl).
- Alkoxy groups include Cj-Qalkoxy and Cj-C 4 alkoxy groups, which have from 1 to 6 or from
- 3-methyipentoxy are representative alkoxy groups.
- alkylthio refers to an alkyl group as attached via a sulfur bridge (i.e., -S-alkyl).
- Alkylthio groups include C r C 6 alkylthio and C r C 4 alky!thio groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively.
- Alkylsulfinyl refers to groups of the formula -(SO)-alky!, in which the sulfur atom is the point of attachment.
- Alkylsulfinyl groups include Cj-Cealkylsulfinyl and Ci-C 4 alkyisulfinyl groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively.
- Alkylsulfonyl refers to groups of the formula -(SO 2 )-alkyl, in which the sulfur atom is the point of attachment. Alkylsuifonyl groups include Ci-C 6 alkylsulfonyl and C r C 4 alkylsulfonyl groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively.
- C 4 alkyi refers to a Cj-C fi alkylsulfonyl that is linked via a single covalent bond or a C r C 4 aIkylene group (i.e., -(C 0 -C 4 alky I)-(S O 2 HC i-C 6 alky I)).
- alkanoyl groups inciude C ⁇ -Cgalkanoyl, Co-Cgalkanoyl and C 2 -
- C 4 alkanoyl groups which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively.
- alkyl ether refers to a linear or branched ether substituent
- Alkyl ether groups include C 2 -C 8 alkyl ether, Q-Qalkyl ether and C 2 -C 4 a!ky! ether groups, which have 2 to 8. 6 or 4 carbon atoms, respectively.
- a C 2 alkyl ether is -CH 2 -O-CH 3 .
- Alkoxycarbonyl groups include C]-Cg, Ci-C 6 and C r C 4 alkoxycarbonyl groups, which have from 1 to 8, 6 or 4 carbon atoms, respectively, in the alkyl portion of the group, "Cj alkoxycarbonyl” refers to
- Alkylamino refers to a secondary or tertiary amine that has the general structure -NH-alkyl or -N(alkyl)(alkyl), wherein each alkyl is selected independently from alkyl, cycloalkyl and (cycloaikyl)alkyl groups.
- groups include, for example, mono- and di-(Ci-C 8 alkyl)amino groups, in which each Cj-Cgalkyi may be the same or different, as well as mono- and di-(C]-C(;alky1)ammo groups and mono- and di-(C r C 4 alkyl)amino groups.
- Alkylaminoalkyl refers to an alkylamino group linked via an alkylene moiety (i.e., a group having the general structure -alkylene— NH-afky! or -aIkylene-N(aikyl)(alkyl)) in which each alkyf is selected independently from alkyl, cycloalkyl and (cycloalkyl)a ⁇ kyl groups.
- Alkylaminoalkyl groups include, for example, mono- and di-(C]-C 3 a]kyl)aminoC,-C6alkyl, and mono- and di-(Q- C 6 alkyl)aminoCi-C 4 alkyl.
- “Mono- or di-(Ci-Qalkyl)aminoC 0 -C 4 alkyi” refers to a mono- or di-(C,- C 6 aikyl)amino group linked via a single covalent bond or a C]-C 4 alkylene group.
- alky as used in the terms “alkylamino” and “alkylaminoalkyl' 1 differs from the definition of "aikyl” used for all other alkyl-containing groups, in the inclusion of cycloalkyl and (cycloalkyl)alkyl groups (e.g., (C 3 -C 7 cycloalkyl)C 0 -C 2 alkyl).
- “Mono- or di-(C r C(;alkyl)aminocarbony]Co-C 4 alkyr refers to an aminocarbonyl group in which one or both hydrogens are replaced with an independently selected Ci-C 6 alkyl group, and which is linked via a single covalent bond or a C r C 4 alkylene group.
- aminosulfonyl refers to a sulfonamide group (i.e., -SO 2 NH 2 ).
- “Mono- or di-(C r C 6 alkyl)aminosulfonylCo-C 4 aikyl” refers to an aminosulfonyl group in which one or both hydrogens are replaced with an independently selected CpQalkyl group, and which is linked via a single covalent bond or a Ci-C 4 alkylene group.
- halogen refers to fluorine, chlorine, bromine or iodine.
- haloalkyl is an alkyl group that is substituted with 1 or more independently chosen halogens (e.g., "Ci-Cghaloaikyl” groups have from 1 to 8 carbon atoms; "Q-Qhaloalkyl” groups have from 1 to 6 carbon atoms).
- haloalkyl groups include, but are not limited to, mono-, di- or tri-fluoromethyl; mono-, di- or tri-chloromethyi; mono-, di-, tri-, tetra- or penta-fiuoroethyl; mono-, di-, tri-, t ⁇ tra- or penta-chloro ⁇ thyl; and 1,2,2,2-tetrafluoro-l-trifluoromethyl-ethyI.
- Typical haloalkyl groups are trifluoromethyl and difluoromethyl.
- a dash (“-") that is not between two letters or numbers is used to indicate a point of attachment for a substituent.
- a "carbocycle” has from 1 to 3 fused, pendant or spiro rings, each of which has only carbon ring members and each of which may, but need not, be bridged by an alkylene moiety.
- a carbocycle that has a single ring contains from 3 to 8 ring members (i.e., C 3 -Cgcarbocycles); rings having from 4 or 5 to 7 ring members (i.e., C 4 -C 7 carbocycles or C 5 -C 7 carbocycles) are recited in certain embodiments.
- Carbocycles comprising fused, pendant or spiro rings typically contain from 9 to 14 ring members.
- Carbocycles may be optionally substituted with a variety of substituents, as indicated.
- a carbocycle may be a cycloalkyl group (i.e., each ring is saturated or partially saturated as described above) or an aryl group (i.e., at least one ring within the group is aromatic).
- Representative aromatic carbocycles are phenyl, ⁇ aphthyi, tetrahydronaphthyl and bi phenyl.
- preferred carbocycles have a single ring, such as phenyl and C 3 - Cscycloalkyl groups.
- carbocycles recited herein are groups (i.e., groups in which a 3- to IO-membered carbocyclic group (which may be cycloalkyl or aryl) is linked via a single covalent bond or a C r C 4 aIkylene group).
- Phenyl groups linked via a single covalent bond or C r C 2 alkylene group are designated phenyiC 0 -C 4 alkyl (e.g., benzyl, l-phenyl-ethyl, 1 -phenyl-propyl and 2-phenyl-ethyl).
- phenyiC 0 -C 4 alkyl e.g., benzyl, l-phenyl-ethyl, 1 -phenyl-propyl and 2-phenyl-ethyl.
- a "phenylC 0 -C4alkoxy” group is a phenyl ring that is linked via an oxygen atom (i.e., phenoxy) or via a Ci-C 4 alkoxy group (e.g., benzyloxy).
- a “heterocycle” (also referred to herein as a “heterocyclic group”) has from 1 to 3 fused, pendant or spiro rings, at least one of which is a heterocyclic ring (i.e., one or more ring atoms is a heteroatom independently chosen from oxygen, sulfur and nitrogen, with the remaining ring atoms being carbon).
- a heterocyclic ring comprises 1 , 2, 3 or 4 heteroatoms; within certain embodiments each heterocyclic ring has 1 or 2 heteroatoms per ring.
- Each heterocyclic ring generally contains from 3 to 8 ring members (rings having from 4 or 5 to 7 ring members are recited in certain embodiments) and heterocycles comprising fused, pendant or spiro rings typically contain from 9 to 14 ring members.
- Certain heterocycles comprise a suifur atom as a ring member; in certain embodiments, the sulfur atom is oxidized to SO or SO 2 .
- Heterocycles may be optionally substituted with a variety of substituents, as indicated.
- Certain heterocycles are 4- to 10-membered and comprise one or two rings; in certain embodiments, such heterocycles are monocyclic (e.g., 4- to 8-rnembered, 5- to 8-membered, 4- to 7-membered, or 5- or 6-membered).
- heterocycles are heteroaiyl groups (i.e., at least one heterocyclic ring within the group is aromatic), such as a 5- to 10-membered heteroaiyl (which may be monocyclic or bicyclic) or a 5- or 6-membered heteroaryl (e.g., thienyl, imidazolyl, pyridy ⁇ or pyrimidyl).
- Other heterocycles are heterocycloalky] groups (i.e., do not comprise an aromatic heterocyclic ring).
- heterocycles may be linked by a single covalent bond or via an alkylene group, as indicated, for example, by the term “(4- to 8-membered heterocycloalkyl)Co-C 4 alkyl.”
- a "substituent,” as used herein, refers to a molecular moiety that is covalently bonded to an atom within a molecule of interest.
- a "ring substituent” may be a moiety such as a halogen, alkyl group, haioalkyl group or other group discussed herein that is covalently bonded to an atom (such as a carbon or nitrogen atom) that is a ring member.
- substitution refers to replacing a hydrogen atom in a molecular structure with a substituent as described above, such that the valence on the designated atom is not exceeded, and such that a chemically stable compound (i.e., a compound that can be isolated, characterized, and tested for biological activity) results from the substitution.
- Groups that are "optionally substituted” are unsubstituted or are substituted by other than hydrogen at one or more available positions, typically 1, 2, 3, 4 or 5 positions, by one or more suitable groups (which may be the same or different).
- Optional substitution is also indicated by the phrase "substituted with from 0 to X substituents," where X is the maximum number of possible substituents.
- Certain optionally substituted groups are substituted with from 0 to 2, 3 or 4 independently selected substituents (i.e., are unsubstituted or substituted with up to the recited maximum number of substituents).
- 1 1 Bi refers to the human Bj bradykinin receptor reported by Menke et al.
- Bi agonist refers to a compound that binds Bi and induces signal transduction mediated by B ⁇ .
- Bi agonists include, for example, bradykinin and kallidin (lysyl-bradykinin), as well as peptide portions or variants of bradykinin or kallidin that bind Bj and retain activity.
- Representative Bj agonists include, but are not limited to, desArg 9 bradykinin and desArg'°kallidin.
- a "Bi antagonist” is a compound that detectably inhibits signal transduction mediated by B 1 . Such inhibition may be determined using the representative calcium mobilization assay provided in Example 7.
- Preferred Bi antagonists have an IC 50 Of 5 ⁇ M or less in this assay, more preferably 2 ⁇ M or less, and still more preferably 1 ⁇ M or less, 500 nM or less, 100 nM or less or IO nM or less.
- the B] antagonist is specific for B 1 (i.e., the IC 50 value in a similar assay performed using the B 2 receptor is greater than 2 ⁇ M and/or the IC 50 ratio (B 2 /Bi) is at least 10, preferably 100, and more preferably at least 1000).
- B] antagonists preferably have minimal agonist activity (i.e., induce an increase in the basal activity of Bi that is less than 5% of the increase that would be induced by one EC 5O of the peptide agonist desArg'°kallidin, and more preferably have no detectable agonist activity within the assay described in Example 7).
- B 1 antagonists for use as described herein are generally non-toxic.
- Bi antagonists include neutral antagonists and inverse agonists.
- a "neutral antagonist" of Bj is a compound that inhibits the activity of B 1 agonist (e.g., desArg'°kallidin) at Bi, but does not significantly change the basal activity of the receptor (i.e., within a calcium mobilization assay as described in Example 7 performed in the absence of agonist, Bj activity is reduced by no more than 10%, more preferably by no more than 5%, and even more preferably by no more than 2%; most preferably, there is no detectable reduction in activity).
- B 1 agonist e.g., desArg'°kallidin
- Bj activity is reduced by no more than 10%, more preferably by no more than 5%, and even more preferably by no more than 2%; most preferably, there is no detectable reduction in activity.
- Neutral antagonists may, but need not, also inhibit the binding of agonist to Bi.
- An "inverse agonist" of B 1 is a compound that reduces the activity of Bi below its basal activity level in the absence of activating concentrations of agonist. Inverse agonists may also inhibit the activity of agonist at Bi, and/or may inhibit binding of B, agonist to B]. The reduction in basal activity of Bi produced by an inverse agonist may be determined from a calcium mobilization assay. such as the assay of Example 7.
- a "therapeutically effective amount” is an amount that, upon administration to a patient, results in a discernible patient benefit (e.g., provides detectable relief from a condition being treated). Such relief may be detected using any appropriate criteria.
- a therapeutically effective amount or dose generally results in a concentration of compound in a body fluid (such as blood. plasma, serum, CSF. synovial fluid, lymph, cellular interstitial fluid, tears or urine) that is sufficient to result in detectable alteration in Br mediated signal transduction (using an assay provided herein).
- the discernible patient benefit may be apparent after administration of a single dose, or may become apparent following repeated administration of the therapeutically effective dose according to a predetermined regimen, depending upon the indication for which the compound is administered. For the treatment of pain, a discernible patient benefit is generally apparent after administration of a single therapeutically effective dose, although further benefit may become apparent following repeated administrations.
- a “patient” is any individual treated with an N-oxide aryl sulfone or sulfoxide as provided herein.
- Patients include humans, as well as other animals such as companion animals (e.g., dogs and cats) and livestock.
- Patients may be experiencing one or more symptoms of a condition responsive to B] modulation or may be free of such symptom(s) ⁇ i.e., treatment may be prophylactic in a patient considered to be at risk for the development of such symptoms).
- the present invention provides N-oxide aryl sulfones and sulfoxides of Formula I that may be used in a variety of contexts, including in the treatment of conditions responsive to B] modulation, as described herein.
- Such compounds may also be used within in vitro assays (e.g., assays for B 1 activity), as probes for detection and localization of Bi and within assays to identify other Bi antagonists.
- Ar is phenyl, naphthyl, pyridyl or pyrimidinyl, each of which is substituted with from 0 to 4 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, C r C 6 alkyL C r C 6 haloalkyl, CpQalkoxy and mono- or
- Ar is phenyl that is substituted with from 0 to 4 substituents independently chosen from Rj (e.g., substituted with 1, 2 or 3 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, C r Q
- each substituent (e.g., I 5 2 or 3 substituents) represented by Ri is independently chosen from halogen, hydroxy, cyano, amino, nitro, Ci-Cealkyl, CVQ,haloalkyl, C r C 6 alkoxy, C]-C 6 haloalkoxy and mono- or di-(C r C 6 alkyl)aminoC 0 -C 4 alkyl.
- Ar is a group of the formula:
- R 4 , R 5 and R 6 are independently chosen from hydrogen, halogen, hydroxy, C r C 6 alkyl, Ci- Qhaloalkyl, C r C 6 alkoxy and CrQhaloalkoxy.
- at least one of R 4 , R 5 and R 5 is not hydrogen; within further such compounds, at least two of R 4 , R 5 and R 6 are not hydrogen; within still further such compounds, R 4 , R 5 and R 6 are independently chosen from halogen, hydroxy, C r C 6 alkyl, Ci-C 6 haloalkyl.
- C r Q,aIkoxy and C r C 6 haioa!koxy e.g., R 5 is methyl or methoxy and R 4 and R 6 are each methyl).
- Y the sum of r and p ranges from 1 to 6 for certain N-oxide aryl sulfones and sulfoxides. Within certain such compounds, p is not zero.
- Certain Y groups satisfy the formula (C H 2 X-O-(CH 2 ) p , which is substituted with from O to 4 substituents independently chosen from amino, hydroxy, cyano, C r C 6 alkyl, C 2 -C 6 alkenyi, C 2 -C 6 aikynyl, d-Qhaloalkyl and substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C 3 -C 6 cycloalkyl. Representative such Y groups include, for example.
- Y is a group of the formula (CH 2 ) r -(CH 2 ) p that is substituted with from O to 4 substituents independently chosen from amino, hydroxy, cyano, C r C 6 alkyl, C 2 -C 6 alkenyl, C 2 -C 6 alkynyl, Ci-Cghaloaikyl and substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C3-C ⁇ ;cycloalkyl.
- Representative such Y groups include, for example, -CH 2 -, -CH 2 -CH 2 -, and -CH 2 -CH 2 -CH 2 -, each of which is optionally substituted as indicated above.
- the variables Rx and Ry of Formula I are, within certain embodiments, independently chosen from hydrogen and C r C 6 aikyl. Within other embodiments, R x and R ⁇ are taken together to form an aromatic group. Representative such aromatic groups include, for example, naphthyl, quinolinyl, phenyl and pyridyl, each of which is optionally substituted as described above. Within further embodiments, R x and Ry are taken together to form an optionally substituted partially or fully saturated ring. Representative such rings include, for example, piperazine, morpholine, thiomorpholine, piperidine and pyrrolidine.
- R x and Ry are taken together to form an optionally substituted 4- to 8-membered heterocycloalkyl.
- X is CH 2 , NR 3 , O, S, SO or SO 2 ; m and q are independently 0, 1 or 2: R 2 represents from 0 to 4 substituents independently chosen from oxo, hydroxy and C
- N-oxide aryl sulfones and sulfoxides of Formula III further satisfy Formula IV or Formula V:
- X is CH 2 in certain compounds; for other compounds X is O.
- R 2 represents from 0 to 4 substituents independently chosen from C
- the variable R c is absent in certain embodiments. It will be apparent that R c may only be absent if R D forms a heteroaryl with R F or a substituent of V (and any heteroaryl so formed comprises V and the tetravalent nitrogen atom as ring members).
- variable "Z" is NR 0 or CR E Rp within certain embodiments of Formulas I-V, It will be apparent that R E , R F and RG are intended to be substituents of the base carbon or nitrogen atom (i.e.,
- R E or R 0 is taken together with R B to form a 4- to 7-membered heterocycloalkyl that is substituted with from 0 to 4 substituents independently chosen from: (i) hydroxy, oxo, cyano and amino; and (ii) C,-C 6 alkyl, C r C 6 alkoxy, mono- or di-(C r C 6 aIkyl)aminoCo-C 4 alkyl, (Cj-Ci ⁇ carbocycle)C 0 -C ⁇ alky] and (4- to 10-membered heteiOcycle)Co-C 4 alkyl; each of which is substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, C r C 6 alkyl, C]-C 6 alkoxy, (C 3 -C i
- s is O, 1 or 2; R D is Ci-Qalkyl; V is absent or Ci- C 4 alkylene; and the remaining variables are as described above.
- s is O, 1 or 2; R D is C r C 6 aIkyl; V is C]-C 4 alkylene; and the remaining variables are as described above.
- R x and R ⁇ are independently hydrogen or Ci-Cealkyl.
- Other compounds of Formula VI or Formula VlI further satisfy Formula VIII or Formula IX, respectively:
- Formula V ⁇ II Formula IX in which m and q are independently O, 1 or 2, such that the sum of m and q is 1, 2 or 3; X is NR 3 , O, S, SO or SO 2 ; R 2 represents from O to 4 substituents independently chosen from oxo, hydroxy and Q- Qalkyl; and the remaining variables are as described for Formula VI. Still further N-oxide aryl sulfones and sulfoxides provided herein satisfy Formula X or
- Formula X Formula Xl in which each s is independently O, 1 or 2.
- R x and R Y are independently hydrogen or Ci-C ⁇ alkyl.
- Other compounds of Formula X or Formula XI further satisfy Formula XII or Formula XIII, respectively: Formula XII Formula XIII in which m and q are independently O, 1 or 2, such that the sum of m and q is 1, 2 or 3;
- X is NR 3 , O, S, SO or SO 2 ;
- R- 2 represents from O to 4 substituents independently chosen from oxo. hydroxy and Ci- C 6 alkyl; and the remaining variables are as described above.
- variable "Z" is CR E R F , and R F is taken together with R D to form a 4- to 7-membered heterocycloalkyl that is substituted with from 0 to 4 substituents independently chosen from: (i) hydroxy, oxo, cyano and amino; and (ii) CVCgalkyl, C.- Qafkoxy, mono- or di- ⁇ C,-C 6 alkyl)aminoCo-C 4 alkyi, (C 3 -Ci 0 carbocycle)C 0 -C 4 a]ky!
- R x and R ⁇ are independently hydrogen or CrQalkyl.
- Other compounds of Formula XIV further satisfy Formula XV:
- Formula XV in which m and q are independently 0. 1 or 2, such that the sum of m and q is 1, 2 or 3; X is NR 3 . O, S, SO or SO 2 ; R 2 represents from 0 to 4 substituents independently chosen from oxo, hydroxy and Ci- Cgalkyi; and the remaining variables are as described above.
- variable "Z" is NR 0 and RQ is hydrogen or CrC 6 alkyl.
- N-oxide aryl sulfones and sulfoxides provided herein include, but are not limited to, those specifically described in the Examples below. It will be apparent that the specific compounds recited herein are representative only, and are not intended to limit the scope of the present invention. Further, as noted above, all compounds of the present invention may be present as a free acid or base or as a pharmaceutically acceptable salt, hydrate or ester.
- compounds provided herein are Bi modulators.
- certain compounds provided herein are specific for B 1 .
- Bj modulator activity may be confirmed using a calcium mobilization assay, such as the assay described in Example 7, herein.
- binding activity of the compounds provided herein to B] may be confi ⁇ ned using the representative assay described in Example 6, herein, or using an assay described by Fox et al.
- Preferred B] modulators exhibit a K 1 within such an assay of 5 micromolar or less, more preferably 2 micromolar or less, 1 micromolar or less, 500 nanomolar or less, 100 nanomolar or less or 10 nanomolar or less.
- Bi modulators In vivo activity of Bi modulators provided herein may be confirmed using any of a variety of animal models including, but not limited to, those described in the following documents (each of which is hereby incorporated by reference for its disclosure of the recited animal model): Wood et al. (2003) J. Med. Chem. 46: 1803-06 - carrageenan-mduced mechanical pressure hyperalgesia; Cottley et al. (2005) Eur. J. Pharmacol, 527:44-51 - thermal antinociception and carrageenan-induced mechanical pressure hypersensitivity;
- compounds provided herein may be evaluated for certain pharmacological properties including, but not limited to, oral bioavailability (preferred compounds are orally bioavailable to an extent allowing for therapeutically effective doses of less than 140 mg/kg, preferably less than 50 mg/kg, more preferably less than 30 mg/kg, even more preferably less than 10 mg/kg, still more preferably less than 1 mg/kg and most preferably less than 0.1 mg/kg), toxicity (a preferred compound is nontoxic when a therapeutically effective amount is administered to a subject), side effects (a preferred compound produces side effects comparable to placebo when a therapeutically effective amount of the compound is administered to a subject), serum protein binding and in vitro and in vivo half-life (a preferred compound exhibits an in vivo half-life allowing for Q. ⁇ .D.
- dosing preferably T.I.D. dosing, more preferably B. LD. dosing, and most preferably once-a- day dosing).
- differential penetration of the blood brain barrier may be desirable. Routine assays that are well known in the art may be used to assess these properties, and identify superior compounds for a particular use. For example, assays used to predict bioavailability include transport across human intestinal cell monolayers, including Caco-2 cell monolayers. Penetration of the blood brain barrier of a compound in humans may be predicted from the brain levels of the compound in laboratory animals given the compound ⁇ e.g., intravenously). Serum protein binding may be predicted from albumin binding assays. Compound half-life is inversely proportional to the frequency of dosage of a compound. In vitro half-iives of compounds may be predicted from assays of microsomal half-life as described herein.
- nontoxic as used herein shall be understood in a relative sense and is intended to refer to any substance that has been approved by the United States Food and Drug Administration (“FDA") for administration to mammals (preferably humans) or, in keeping with established criteria, is susceptible to approval by the FDA for administration to mammals (preferably humans).
- FDA United States Food and Drug Administration
- a highly preferred nontoxic compound generally satisfies one or more of the following criteria: (1) does not substantially inhibit cellular ATP production; (2) does not significantly prolong heart QT intervals; (3) does not cause substantial liver enlargement, or (4) does not cause substantial release of liver enzymes.
- a compound that does not substantially inhibit cellular ATP production is a compound that satisfies the criteria set forth in Example 8, herein.
- cells treated as described in Example 8 with 100 ⁇ M of such a compound exhibit ATP levels that are at least 50% of the ATP levels detected in untreated cells.
- such cells exhibit
- ATP levels that are at least 80% of the ATP levels detected in untreated cells.
- a compound that does not significantly prolong heart QT intervals is a compound that does not result in a statistically significant prolongation of heart QT intervals (as determined by electrocardiography) in guinea pigs, minipigs or dogs upon administration of a dose that yields a serum concentration equal to the EC 50 or IC 50 for the compound.
- a dose of 0.01 , 0.05, 0.1, 0.5, 1 , 5, 10, 40 or 50 mg/kg administered parenterally or orally does not result in a statistically significant prolongation of heart QT intervals.
- statically significant results varying from control at the p ⁇ 0.1 level or more preferably at the p ⁇ 0.05 level of significance as measured using a standard parametric assay of statistical significance such as a student's T test.
- a compound does not cause substantial liver enlargement if daily treatment of laboratory rodents (e.g., mice or rats) for 5-10 days with a dose that yields a serum concentration equal to the EC 50 or IC 50 for the compound results in an increase in liver to body weight ratio that is no more than 100% over matched controls. In more highly preferred embodiments, such doses do not cause liver enlargement of more than 75% or 50% over matched controls.
- non-rodent mammals e.g., dogs
- such doses should not result in an increase of liver to body weight ratio of more than 50%, preferably not more than 25%. and more preferably not more than 10% over matched untreated controls.
- Preferred doses within such assays include 0.01, 0.05. 0.1 , 0.5, 1, 5, 10, 40 or 50 mg/kg administered parenterally or orally.
- a compound does not promote substantial release of liver enzymes if administration of twice the minimum dose that yields a serum concentration equal to the EC 50 or IC 5 O for the compound does not elevate serum levels of ALT, LDH or AST in laboratory rodents by more than 100% over matched mock-treated controls. In more highly preferred embodiments, such doses do not elevate such serum levels by more than 75% or 50% over matched controls.
- a compound does not promote substantial release of liver enzymes if, in an in vitro hepatocyte assay, concentrations (in culture media or other such solutions that are contacted and incubated with hepatocytes in vitro) that are equal to the EC 50 or IC 50 for the compound do not cause detectable release of any of such liver enzymes into culture medium above baseline levels seen in media from matched mock-treated control cells. In more highly preferred embodiments, there is no detectable release of any of such liver enzymes into culture medium above baseline levels when such compound concentrations are five-fold, and preferably ten-fold the EC 50 or IC 50 for the compound.
- certain preferred compounds do not inhibit or induce microsomal cytochrome P450 enzyme activities, such as CYP 1A2 activity, CYP2A6 activity, CYP2C9 activity, CYP2C19 activity, CYP2D6 activity, CYP2EI activity or CYP3A4 activity at a concentration equal to the EC 50 or IC 50 for the compound.
- microsomal cytochrome P450 enzyme activities such as CYP 1A2 activity, CYP2A6 activity, CYP2C9 activity, CYP2C19 activity, CYP2D6 activity, CYP2EI activity or CYP3A4 activity at a concentration equal to the EC 50 or IC 50 for the compound.
- Certain preferred compounds are not clastogenic (e.g., as determined using a mouse erythrocyte precursor cell micronucieus assay, an Ames micronucleus assay, a spiral micronucleus assay or the like) at a concentration equal the EC 50 or IC 50 for the compound.
- certain preferred compounds do not induce sister chromatid exchange (e.g., in Chinese hamster ovary cells) at such concentrations.
- compounds provided herein may be isotopically-Iabeled or radiolabeled.
- such compounds may have one or more atoms replaced by an atom of the same element having an atomic mass or mass number different from the atomic mass or mass number usually found in nature.
- isotopes that can be present in the compounds provided herein include isotopes of hydrogen, carbon, nitrogen, oxygen, phosphorous, fluorine and chlorine, such as 2 H, 3 H. 11 C, 13 C. 14 C, 15 N, 18 O, 17 O, 31 P, 32 P, 35 S, 18 F and 36 Cl.
- substitution with heavy isotopes such as deuterium can afford certain therapeutic advantages resulting from greater metabolic stability, for example increased in vivo half-life or reduced dosage requirements and, hence, may be preferred in some circumstances.
- Suitable protecting groups and methodology for protection and deprotection such as those described in Protecting Groups in Organic Synthesis by T. Greene, are well known. Compounds and intermediates requiring protection/deprotection will be readily apparent.
- a compound provided herein may contain one or more asymmetric carbon atoms, so that the compound can exist in different stereo isomeric forms.
- Such forms can be, for example, racemates or optically active forms.
- All stereoisomers are encompassed by the present invention. Nonetheless, it may be desirable to obtain single enantiomers ⁇ i.e., optically active forms).
- Standard methods for preparing single enantiomers include asymmetric synthesis and resolution of the racemates. Resolution of the racemates can be accomplished, for example, by conventional methods such as crystallization in the presence of a resolving agent, or chromatography using, for example a chiral HPLC column.
- Compounds may be radiolabeled by carrying out their synthesis using precursors comprising at least one atom that is a radioisotope.
- Each radioisotope is preferably carbon (e.g., 14 C), hydrogen (e.g., 3 H), sulfur (e.g., 3 ⁇ S) or iodine (e.g., 125 I).
- Tritium labeled compounds may also be prepared catalytically via platinum-catalyzed exchange in tritiated acetic acid, acid-catalyzed exchange in tritiated trifluoroacetic acid, or heterogeneous-catalyzed exchange with tritium gas using che compound as substrate.
- certain precursors may be subjected to tritium-halogen exchange with tritium gas, tritium gas reduction of unsaturated bonds, or reduction using sodium borotritide, as appropriate.
- Preparation of radiolabeled compounds may be conveniently performed by a radioisotope supplier specializing in custom synthesis of radiolabeled probe compounds.
- compositions comprising one or more N- oxide ary! sulfones and sulfoxides provided herein, together with at least one physiologically acceptable carrier or excipient.
- Pharmaceutical compositions may comprise, for example, one or more of water, buffers (e.g., neutral buffered saline or phosphate buffered saline), ethanoi, mineral oil, vegetable oil, dimethylsulfoxide, carbohydrates (e.g., glucose, mannose, sucrose or dextrans), rnannitol, proteins, adjuvants, polypeptides or amino acids such as glycine, antioxidants, chelating agents such as EDTA or glutathione and/or preservatives.
- other active ingredients may (but need not) be included in the pharmaceutical compositions provided herein.
- compositions may be formulated for any appropriate manner of administration, including, for example, topical, oral (including, but not limited to, sublingual), nasa ⁇ , rectal or parenteral administration.
- parenteral as used herein includes subcutaneous, intradermal, intravascular (e.g., intravenous), intramuscular, spina!, intracranial, intrathecal and intraperitoneal injection, as well as any similar injection or infusion technique.
- compositions suitable for oral use are preferred. Such compositions include, for example, tablets, troches, lozenges, aqueous or oily suspensions, dispersible powders or granules, emulsion, hard or soft capsules, or syrups or elixirs.
- compositions of the present invention may be formulated as a lyophilizate.
- Compositions intended for oral use may further comprise one or more components such as sweetening agents, flavoring agents, coloring agents and/or preserving agents in order to provide appealing and palatable preparations.
- Tablets contain the active ingredient in admixture with physiologically acceptable excipients that are suitable for the manufacture of tablets.
- excipients include, for example, inert diluents (e.g., calcium carbonate, sodium carbonate, lactose, calcium phosphate or sodium phosphate), granulating and disintegrating agents (e.g., corn starch or alginic acid), binding agents (e.g., starch, gelatin or acacia) and lubricating agents (e.g., magnesium stearate, stearic acid or talc).
- inert diluents e.g., calcium carbonate, sodium carbonate, lactose, calcium phosphate or sodium phosphate
- granulating and disintegrating agents e.g., corn starch or
- the tablets may be uncoated or they may be coated by known techniques to delay disintegration and absorption in the gastrointestinal tract and thereby provide a sustained action over a longer period.
- a time delay material such as glyceryl monosterate or glyceryl distearate may be employed.
- Formulations for oral use may also be presented as hard gelatin capsules wherein the active ingredient is mixed with an inert solid diluent (e.g., calcium carbonate, calcium phosphate or kaolin), or as soft gelatin capsules wherein the active ingredient is mixed with water or an oil medium (e.g., peanut oil, liquid paraffin or olive oil).
- an inert solid diluent e.g., calcium carbonate, calcium phosphate or kaolin
- an oil medium e.g., peanut oil, liquid paraffin or olive oil
- Aqueous suspensions contain the active material(s) in admixture with excipients suitable for the manufacture of aqueous suspensions.
- excipients include suspending agents (e.g., sodium carboxymethylcellulose, methylceliulose, hydropropylmethylcellulose, sodium alginate, polyvinylpyrrolidone, gum tragacanth and gum acacia); and dispersing or wetting agents (e.g., naturally-occurring phosphatides such as lecithin, condensation products of an alkylene oxide with fatty acids such as polyoxyethylene stearate, condensation products of ethylene oxide with long chain aliphatic alcohols such as heptadecaethyleneoxycetanol, condensation products of ethylene oxide with partial esters derived from fatty acids and a hexitoi such as polyoxyethylene sorbitol monooleate, or condensation products of ethylene oxide with partial esters derived from fatty acids and hexitoi anhydrides such as polyethylene sorbitan
- Aqueous suspensions may also comprise one or more preservatives, such as ethyl or n-propyl p-hydroxybenzoate, one or more coloring agents, one or more flavoring agents, and one or more sweetening agents, such as sucrose or saccharin.
- Oily suspensions may be formulated by suspending the active ingredient(s) in a vegetable oil ⁇ e.g., arachis oil, olive oil, sesame oil or coconut oil) or in a mineral oil such as liquid paraffin.
- the oily suspensions may contain a thickening agent such as beeswax, hard paraffin or cetyl alcohol. Sweetening agents such as those set forth above, and/or flavoring agents may be added to provide palatable oral preparations.
- Such suspensions may be preserved by the addition of an anti-oxidant such as ascorbic acid.
- Dispersible powders and granules suitable for preparation of an aqueous suspension by the addition of water provide the active ingredient in admixture with a dispersing or wetting agent, suspending agent and one or more preservatives.
- a dispersing or wetting agent e.g., glycerol, glycerol, glycerol, glycerol, glycerol, glycerol, glycerin, glycerin, glycerin, glycerin, glycerin, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, sorbitol, glycerol, glycerol, glycerol, glycerol, glycerol, glycerol, glycerol, glycerol, glycerol
- compositions may also be formulated as oil-in-water emulsions.
- the oily phase may be a vegetable oil ⁇ e.g., olive oil or arachis oil), a mineral oil (e.g., liquid paraffin) or a mixture thereof.
- Suitable emulsifying agents include naturally-occurring gums (e.g., gum acacia or gum tragacanth), naturally-occurring phosphatides (e.g., soy bean lecithin, and esters or partial esters derived from fatty acids and hexitoi), anhydrides (e.g., sorbitan monoleate) and condensation products of partial esters derived from fatty acids and hexitoi with ethylene oxide (e.g., polyoxyethylene sorbitan monoleate).
- An emulsion may also comprise one or more sweetening and/or flavoring agents.
- Syrups and elixirs may be formulated with sweetening agents, such as glycerol, propylene glycol, sorbitol or sucrose. Such formulations may also comprise one or more demulcents, preservatives, flavoring agents and/or coloring agents.
- sweetening agents such as glycerol, propylene glycol, sorbitol or sucrose.
- Such formulations may also comprise one or more demulcents, preservatives, flavoring agents and/or coloring agents.
- Formulations for topical administration typically comprise a topical vehicle combined with active agent(s), with or without additional optional components.
- Suitable topical vehicles and additional components are well known in the art, and it will be apparent that the choice of a vehicle will depend on the particular physical form and mode of delivery.
- Topical vehicles include water; organic solvents such as alcohols (e.g., ethanol or isopropyl alcohol) or glycerin; glycols (e.g., butylene, isoprene or propylene glycol); aliphatic alcohols (e.g., lanolin); mixtures of water and organic solvents and mixtures of organic solvents such as alcohol and glycerin; lipid-based materials such as fatty acids, acylglycerols (including oils, such as mineral oil, and fats of natural or synthetic origin), phosphoglycerides, sphingolipids and waxes; protein-based materials such as collagen and gelatin; silicone-based materials (both non-volatile and volatile): and hydrocarbon-based materials such as m ⁇ crosponges and polymer matrices.
- organic solvents such as alcohols (e.g., ethanol or isopropyl alcohol) or glycerin
- glycols e.g., butylene, iso
- a composition may further include one or more components adapted to improve the stability or effectiveness of the applied formulation, such as stabilizing agents, suspending agents, emulsifying agents, viscosity adjusters, gelling agents, preservatives, antioxidants, skin penetration enhancers, moisturizers and sustained release materials.
- stabilizing agents such as hydroxymethylcellulose or gelatin-microcapsules, liposomes, albumin microspheres, microemuisions, nanoparticles or nanocapsules.
- a topical formulation may be prepared in a variety of physical forms including, for example, sofids, pastes, creams, foams, lotions, gels, powders, aqueous liquids and emulsions.
- Typical modes of delivery for topical compositions include application using the fingers; application using a physical applicator such as a cloth, tissue, swab, stick or brush; spraying (including mist, aerosol or foam spraying); dropper application; sprinkling; soaking; and rinsing. Controlled release vehicles can also be used.
- a pharmaceutical composition may be prepared as a sterile injectible aqueous or oleaginous suspension.
- the compound(s) provided herein can either be suspended or dissolved in the vehicle.
- a composition may be formulated according to the known art using suitable dispersing, wetting and/or suspending agents such as those mentioned above.
- suitable dispersing, wetting and/or suspending agents such as those mentioned above.
- the acceptable vehicles and solvents that may be employed are water, 1,3-butanedioi, Ringer's solution and isotonic sodium chloride solution.
- sterile, fixed oils may be employed as a solvent or suspending medium.
- any bland fixed oil may be employed, including synthetic mono- or diglycerides.
- fatty acids such as oleic acid find use in the preparation of injectible compositions, and adjuvants such as local anesthetics, preservatives and/or buffering agents can be dissolved in the vehicle.
- Compounds may also be formulated as suppositories (e g., for rectal administration). Such compositions can be prepared by mixing the drug with a suitable non-irritating excipient that is solid at ordinary temperatures but liquid at the rectal temperature and will therefore melt in the rectum to release the drug. Suitable excipients include, for example, cocoa butter and polyethylene glycols.
- compositions may be formulated for release at a pre-determined rate.
- Instantaneous release may be achieved, for example, via sublingual administration (i.e., administration by mouth in such a way that the active ingredient(s) are rapidly absorbed via the blood vessels under the tongue rather than via the digestive tract).
- Controlled release formulations i.e., formulations such as a capsule, tablet or coated tablet that slows and/or delays release of active ingredient(s) foliowing administration
- a controlled release formulation comprises a matrix and/or coating that delays disintegration and absorption in the gastrointestinal tract (or implantation site) and thereby provides a delayed action or a sustained action over a longer period.
- One type of control led-release formulation is a sustained-release formulation, in which at least one active ingredient is continuously released over a period of time at a constant rate.
- the therapeutic agent is released at such a rate that blood (e.g., plasma) concentrations are maintained within the therapeutic range, but below toxic levels, over a period of time that is at least 4 hours, preferably at least 8 hours, and more preferably at least 12 hours.
- Such formulations may generally be prepared using well known technology and administered by, for example, oral, rectal or subcutaneous implantation, or by implantation at the desired target site.
- Carriers for use within such formulations are biocompatible, and may also be biodegradable; preferably the formulation provides a relatively constant level of modulator release.
- the amount of modulator contained within a sustained release formulation depends upon, for example, the site of implantation, the rate and expected duration of release and the nature of the condition to be treated or prevented.
- Controlled release may be achieved by combining the active ingredient(s) with a matrix material that itself alters release rate and/or througii the use of a controlled-release coating.
- the release rate can be varied using methods well known in the art, including (a) varying the thickness or composition of coating, (b) altering the amount or manner of addition of plasticizer in a coating, (c) including additional ingredients, such as release-modifying agents, (d) altering the composition, particle size or particle shape of the matrix, and (e) providing one or more passageways through the coating.
- the amount of modulator contained within a sustained release formulation depends upon, for example, the method of administration (e.g., the site of implantation), the rate and expected duration of release and the nature of the condition to be treated or prevented.
- the matrix material which itself may or may not serve a controlled-release function, is generally any material that supports the active ingredient(s).
- a time delay material such as glyceryl monosterate or glyceryl distearate may be employed.
- Active ingredient(s) may be combined with matrix material prior to formation of the dosage form (e.g., a tablet).
- active ingredient(s) may be coated on the surface of a particle, granule, sphere, microsphere, bead or pellet that comprises the matrix material. Such coating may be achieved by conventional means, such as by dissolving the active ingredient(s) in water or other suitable solvent and spraying.
- additional ingredients are added prior to coating (e.g., to assist binding of the active ingredient(s) to the matrix material or to color the solution).
- the matrix may then be coated with a barrier agent prior to application of controlled-release coating. Multiple coated matrix units may, if desired, be encapsulated to generate the final dosage fo ⁇ n.
- a controlled release is achieved through the use of a controlled release coating (i.e., a coating that permits release of active ingredient(s) at a controlled rate in aqueous medium).
- the controlled release coating should be a strong, continuous film that is smooth, capable of supporting pigments and other additives, non-toxic, inert and tack-free.
- Coatings that regulate release of the modulator include pH-independent coatings, pH-dependent coatings (which may be used to release modulator in the stomach) and enteric coatings (which allow the formulation to pass intact through the stomach and into the small intestine, where the coating dissolves and the contents are absorbed by the body).
- pH dependent coatings include, for example, shellac, cellulose acetate phthalate, polyvinyl acetate phthalate, hydroxypropylmethylcellulose phthalate, methacrylic acid ester copolymers and zein.
- the coating is a hydrophobic material, preferably used in an amount effective to slow the hydration of the gelling agent following administration.
- Suitable hydrophobic materials include alkyl celluloses (e.g., ethylcellulose or carboxymethylcellulose), cellulose ethers, cellulose esters, acrylic polymers (e.g., po!y(acrylic acid), poiy(methacrylic acid), acrylic acid and methacrylic acid copolymers, methyl methacrylate copolymers, ethoxy ethyl methacrylates, cyanoethyl methacrylate, methacrylic acid alkamide copolymer, poly(methyl methacrylate), polyacrylamide, ammonio methacrylate copolymers, aminoalkyi methacrylate copolymer, ⁇ oly(methacrylic acid anhydride) and glycidyl methacrylate copolymers) and mixtures of the foregoing.
- AQUACOAT® FMC Corp., Philadelphia, PA
- SURELEASE® Colorcon, Inc., West Point
- acrylic polymers include, for example, the various EUDRAGIT® (Rohm America,
- Piscataway, NJ Polymers, which may be used singly or in combination depending on the desired release profile, according to the manufacturer's instructions.
- Suitable plasticizers for alkyl celluloses include, for example, dibutyi sebacate, diethyl phthalate, triethyl citrate, tributyl citrate and triacetin.
- Suitable plasticizers for acrylic polymers include, for example, citric acid esters such as triethyl citrate and tributyl citrate, dibutyi phthalate, polyethylene glycols, propylene glycol, diethyl phthalate, castor oii and triacetin.
- Controlled-release coatings are generally applied using conventional techniques, such as by spraying in the form of an aqueous dispersion.
- the coating may comprise pores or channels to facilitate release of active ingredient. Pores and channels may be generated by well known methods, including the addition of organic or inorganic material that is dissolved, extracted or leached from the coating in the environment of use.
- pore-forming materials include hydrophilic polymers, such as hydroxyalkylcelluloses (e g., hydroxypropylmethylcellulose), cellulose ethers, synthetic water-soluble polymers (e.g., polyvinylpyrrolidone, cross-linked polyvinylpyrrolidone and polyethylene oxide), water-soluble polydextrose, saccharides and polysaccharides and alkali metal salts.
- a controlled release coating may include one or more orifices, which may be formed my methods such as those described in US Patent Nos. 3,845,770; 4,034,758; 4,077,407; 4,088,864; 4,783,337 and 5,071,607. Controlled-release may also be achieved through the use of tran sdermal patches, using conventional technology ⁇ see, e.g., US Patent No. 4,668,232).
- controlled release formulations and components thereof, may be found, for example, in US Patent Nos. 4,572,833; 4,587,1 17; 4,606,909; 4,610,870; 4,684,516; 4,777,049;
- a compound provided herein may be conveniently added to food or drinking water ⁇ e.g., for administration to non-human animals including companion animals (such as dogs and cats) and livestock).
- Animal feed and drinking water compositions may be formulated so that the animal takes in an appropriate quantity of the composition along with its diet. It may also be convenient to present the composition as a premix for addition to feed or drinking water.
- N-oxide aryl sulfones and sulfoxides provided herein are generally administered in a therapeutically effective amount.
- Preferred systemic doses are no higher than 50 mg per kilogram of body weight per day ⁇ e.g., ranging from about 0.001 mg to about 50 mg per kilogram of body weight per day), with oral doses generally being about 5-20 fold higher than intravenous doses ⁇ e.g., ranging from 0.01 to 40 mg per kilogram of body weight per day).
- the amount of active ingredient that may be combined with the carrier materials to produce a single dosage unit will vaiy depending, for example, upon the patient being treated and the particular mode of administration. Dosage units will generally contain from about 10 ⁇ g to about 500 mg of an active ingredient. Optimal dosages may be established using routine testing, and procedures that are well known in the art.
- compositions provided herein may, but need not, further comprise one or more additional pharmaceutical agents, such as an anti-inflammatory agent or analgesic.
- Anti-inflammatory agents include, for example, non-steroidal anti-inflammatory drugs
- NSAIDs non-specific and cyclooxygenase-2 (COX-2) specific cyciooxgenase enzyme inhibitors, gold compounds, corticosteroids, methotrexate, leflunomide, cyclosporine A, IM gold, minocycline, azathioprine, tumor necrosis factor (TNF) receptor antagonists, soluble TNF alpha receptor
- IL-I interleukin- 1 receptor antagonists ⁇ e.g., anakinra or IL-I trap), IL- 18 binding protein, CTLA4-
- Ig ⁇ e.g., abatacept anti-human IL-6 receptor monoclonal antibody ⁇ e.g., tocilizumab
- LFA-3-lg fusion proteins ⁇ e.g., alefacept
- LFA-I antagonists anti-VLA4 monoantibody ⁇ e.g., natalizumab
- anti-CDl la monoclonal antibody anti-CD20 monoclonal antibody ⁇ e.g., rituximab
- anti-IL-12 monoclonal antibody anti-IL-15 monoclonal antibody
- CDP 484, CDP 870 chemokine receptor antagonists, selective iNOS inhibitors, p38 kinase inhibitors, integrin antagonists, angiogenesis inhibitors, and TMI-! dual inhibitors.
- Further anti-inflammatory agents include meloxicam, rofecoxib, celecoxib, etoricoxib, parecoxib, valciecoxib and tilicoxib.
- NSAIDs include, but are not limited to, ibuprofen, flurbiprofen, naproxen or naproxen sodium, diclofenac, combinations of diclofenac sodium and misoprostol, sulindac, oxaprozin, diflunisal, piroxicam, indomethacin, etodolac, fenoprofen calcium, ketoprofen, sodium nabumetone, sulfasalazine, tolmetin sodium, and hydroxychloroquine.
- One class of NSAIDs consists of compounds that inhibit eye 1 ⁇ oxygenase (COX) enzymes; such compounds include celecoxib and rofecoxib.
- NSAlDs further include salicylates such as acetylsalicylic acid or aspirin, sodium salicylate, choline and magnesium salicylates, and salsalate, as well as corticosteroids such as cortisone, dexamethasone, methylprednisolone, prednisolone, prednisolone sodium phosphate, and prednisone.
- salicylates such as acetylsalicylic acid or aspirin
- sodium salicylate sodium salicylate
- choline and magnesium salicylates sodium salicylate
- salsalate sodium salicylate
- corticosteroids such as cortisone, dexamethasone, methylprednisolone, prednisolone, prednisolone sodium phosphate, and prednisone.
- analgesics for use in combination with B] modulators are also antiinflammatory agents, and are listed above.
- Other such medications are analgesic agents, including narcotic agents which typically act at one or more opioid receptor subtypes (e.g., ⁇ , K and/or ⁇ ), preferably as agonists or partial agonists.
- opioid receptor subtypes e.g., ⁇ , K and/or ⁇
- Such agents include opiates, opiate derivatives and opioids, as well as pharmaceutically acceptable salts and hydrates thereof.
- narcotic analgesics include, within preferred embodiments, alfentanil, a ⁇ phaprodine, anileridine, bezitramide, buprenorphine, butorphanol, codeine, diacetyidihydromorphine, diacetylmorphine, dihydrocodeine, diphenoxylate, ethylmorphine, fentanyl, heroin, hydrocodone, hydromorphone, isomethadone, levomethorphan, levorphane, levorphanol, meperidine, metazocine, methadone, methorphan, metopon, morphine, nalbuphine, opium extracts, opium fluid extracts, powdered opium, granulated opium, raw opium, tincture of opium, oxycodone, oxymorphone, paregoric, pentazocine, pethidine, phenazocine, piminodine, propoxyphene, racemeth
- narcotic analgesic agents include acetorphine, acetyldihydrocodeine, acetylmethadol, allylprodine, alphracetylmethadol, alphameprodine, alphamethadol, benzethidine, benzyimo ⁇ hine, betacetylmethadol, betameprodine, betamethadol, betaprodine, clonitazene, codeine methylbromide, codeine-N-oxide, cypreno ⁇ hine, desomorphine, dextromoramide, diampromide, diethylthiambutene, dihydromorphine, dimenoxadol, dimepheptanol, dimethylthiamubutene, dioxaphetyl buryrate.
- naloxone naltyhexone, nicocodeine, ⁇ icomo ⁇ hine, noracymethadol, norlevo ⁇ hanol, normethadone, normo ⁇ hine, no ⁇ ipanone, pentazocaine, phenadoxone, phenampromide, phenomo ⁇ han, phenoperidine, piritramide, pholcodine, proheptazoine, properidine, prop Iran, racemoramide, thebacon, trimeperidine and the pharmaceutically acceptable salts and hydrates thereof.
- analgesic agents include, for example acetaminophen
- NR2B antagonists capsaicin receptor antagonists
- anti-migraine agents such as oxcarbazepine and carbamazepine
- antidepressants such as TCAs, SSRIs, SNRIs, substance P antagonists, etc.
- spinal blocks pentazocine/naloxone; meperidine; levorphanol; buprenorphine; hydromorphone; fentanyl; sufentanyi; oxycodone; oxycodone/acetaminophen, nalbuphine and oxymorphone.
- analgesic agents include CB2-receptor agonists, such as AM 1241, capsaicin receptor antagonists and compounds that bind to the ⁇ 2 ⁇ subunit of voltage-gated calcium channels, such as gabapentin and pregabalin.
- compositions may be packaged for treating conditions responsive to B] modulation ⁇ e.g., treatment of pain, inflammation or other disorder(s) recited herein).
- Packaged pharmaceutical preparations generally comprise a container holding a therapeutically effective amount of a pharmaceutical composition as described above and instructions (e.g., labeling) indicating that the composition is to be used for treating a condition responsive to B s modulation in a patient (e.g., pain or other disorder as indicated herein).
- a packaged pharmaceutical preparation comprises one or more N-oxide aryl sulfone or sulfoxide provided herein and one or more additional agents in the same package, either in separate containers within the package or in the same container (i.e., as a mixture). Preferred mixtures are formulated for oral administration (e.g., as pills, capsules, tablets or the like).
- the package comprises a label bearing indicia indicating that the components are to be taken together for the treatment of pain.
- the present invention provides methods for treating a condition responsive to B] modulation in a patient.
- the patient may be afflicted with such a condition, or may be free of symptoms but considered at risk for developing such a condition.
- a condition is "responsive to B 1 modulation' 1 if the condition or symptom(s) thereof are alleviated, attenuated, delayed or otherwise improved by modulation of B s activity.
- such methods comprise administering to the patient a therapeutically effective amount of at least one N-oxide aiyl suifone or sulfoxide as provided herein.
- Conditions responsive to B 1 modulation include, for example pain; inflammation including neuroinflammation (such as atherosclerosis), inflammation associated with airway diseases (e.g., asthma, including allergic asthma, exercise-induced bronchoconstriction, occupational asthma, and other non-allergic asthmas), and inflammatory skin disorders (e.g., psoriasis and eczema)); respiratory disorders including bronchoconstriction, asthma, chronic obstructive pulmonary disease (e.g., emphysema), chronic cough (including ACE-inhibitor cough), adult respiratory distress syndrome, bronchitis, pneumonia, allergic rhinitis and vasomotor rhinitis; vascular edema (including diabetes- related vascular disease); and epilepsy.
- neuroinflammation such as atherosclerosis
- airway diseases e.g., asthma, including allergic asthma, exercise-induced bronchoconstriction, occupational asthma, and other non-allergic asthmas
- inflammatory skin disorders e.g.
- diabetes e.g., type II or non insulin dependent, as well as diabetic vasculopathy, diabetic neuropathy, diabetic retinopathy, post capillary resistance and symptoms associated with insulitis
- seizure disorders e.g., epilepsy
- multiple sclerosis liver disease
- cardiovascular disorders e.g., atherosclerosis, congestive heart failure and myocardial infarction
- neurodegenerative diseases e.g., Alzheimer's disease and Parkinson's disease
- rheumatoid arthritis infection, cancer, crania!
- Bj modulators may also be used as smooth muscle relaxants for treating spasms of the gastrointestinal tract of uterus.
- the condition responsive to B] modulation is pain or inflammation.
- Pain that may be treated using the Bi modulators provided herein may be chronic or acute and includes, but is not limited to, peripheral nerve-mediated pain (especially neuropathic pain, such as pain due to diabetes, postherpetic neuralgia, nerve injury, vulvodynia, root avulsions, painful traumatic neuropathy and painful polyneuropathy).
- peripheral nerve-mediated pain especially neuropathic pain, such as pain due to diabetes, postherpetic neuralgia, nerve injury, vulvodynia, root avulsions, painful traumatic neuropathy and painful polyneuropathy).
- visceral pain e.g., pancreatitis, interstitial cystitis and renal colic
- persistent hyperalgesia inflammatory pain, repetitive motion pain, carpel tunnel syndrome, perioperative pain, algesia, oral neuropathic pain, toothache (dental pain), denture pain, ocular pain, postherpetic neuralgia, diabetic neuropathy, chemotherapy- induced neuropathy, reflex sympathetic dystrophy, trigeminal neuralgia, bone and joint pain (e.g., pain associated with osteoarthritis), rheumatoid arthritis, myofascial pain (e.g., muscular injury and fibromyalgia), Guillain-Barre syndrome, meralgia paresthetica, burning-mouth syndrome and/or pain associated with nerve and root damage, including as pain associated with peripheral nerve disorders (e.g., nerve entrapment and brachial plexus avulsions, amputation,
- peripheral nerve disorders e.g., nerve entrapment and
- Additional neuropathic pain conditions include causalgia (reflex sympathetic dystrophy - RSD, secondary to injury of a peripheral nerve), neuritis (including, for example, sciatic neuritis, peripheral neuritis, polyneuritis, optic neuritis, postfebrile neuritis, migrating neuritis, segmental neuritis and Gombault's neuritis), neuronitis, neuralgias (e.g., those mentioned above, cervicobrachial neuralgia, cranial neuralgia, geniculate neuralgia, glossopharyngial neuralgia, migranous neuralgia, idiopathic neuralgia, intercostals neuralgia, mammary neuralgia, mandibular joint neuralgia, Morton's neuralgia, nasociliary neuralgia, occipital neuralgia, red neuralgia, Binder's neuralgia, splenopalatine neuralgi
- Headache including headaches involving peripheral nerve activity may also be treated as described herein.
- Such headache pain includes, for example, sinus, cluster (i.e., migranous neuralgia) and tension headaches, migraine, temporomandibular pain and maxillary sinus pain.
- migraine headaches may be prevented by administration of a compound provided herein as soon as a pre-migrainous aura is experienced by the patient.
- Charcot's pains intestinal gas pains, ear pain, heart pain, muscle pain, eye pain, orofacial pain (e.g., odontalgia), abdominal pain, gynaec ⁇ logical pain (e.g., menstrual pain, dysmenorrhoea, pain associated with cystitis, labor pain, chronic pelvic pain, chronic prostitis and endometriosis), acute and chronic back pain (e.g., lower back pain), gout, scar pain, hemorrhoidal pain, dyspeptic pains, pain associated with angina, nerve root pain, "non-painful" neuropathies, complex regional pain syndrome, nomotopic pain and heterotopic pain — including pain associated with carcinoma, often referred to as cancer pain (e.g., in patients with bone cancer), pain (and inflammation) associated with venom exposure (e.g., due to snake bite, spider bite, or insect sting) and trauma associated pain (e.g., post-
- pain treated with B] modulators provided herein is inflammatory pain, acute pain, dental pain, back pain, surgical pain, headache, neuropathic pain or pain from osteoarthritis or trauma.
- compounds provided herein may be administered alone or in combination with one or more additional agents that are suitable for treating the disorder of interest.
- the compound(s) and additional agent(s) may be present in the same pharmaceutical composition, or may be administered separately in either order.
- Representative anti-inflammatory agents and analgesics for use in combination therapy include those indicated above.
- modulators provided herein may be used within combination therapy for the treatment of conditions involving pain and/or inflammatory components.
- Such conditions include, for example, autoimmune disorders and pathologic autoimmune responses known to have an inflammatory component including, but not limited to, arthritis (especially rheumatoid arthritis), psoriasis, Crohn's disease, lupus erythematosus, irritable bowel syndrome, tissue graft rejection, and hyperacute rejection of transplanted organs.
- Other such conditions include trauma (e.g., injury to the head or spinal cord), cardio- and cerebro-vascular disease and certain infectious diseases.
- a B] modulator is administered to a patient along with an additional analgesic and/or anti-inflammatory agent.
- the B; modulator and additional analgesic and/or anti- inflammatory agent may be present in the same pha ⁇ naceutical composition, or may be administered separately in either order.
- Administration to the patient can be by way of any means discussed above, including oral, topical, nasal or transdermal administration, or intravenous, intramuscular, subcutaneous, intrathecal, epidural, intracerebroventricular or like injection.
- Oral administration is preferred in certain embodiments (e.g., formulated as pills, capsules, tablets or the like).
- Treatment regimens may vary depending on the compound used and the particular condition to be treated. In genera!, a dosage regimen of 4 times daily or less is preferred, with 1 or 2 times daily particularly preferred. It will be understood, however, that the specific dose and treatment regimen for any particular patient will depend upon a variety of factors including the activity of the specific compound employed, the age, body weight, general health, sex and diet of the patient, the time of administration, the route of administration, the rate of excretion, any drug combination and the severity of the particular disease undergoing therapy. Dosages are generally as described above; in general, the use of the minimum dose sufficient to provide effective therapy is preferred. Patients may generally be monitored for therapeutic effectiveness using medical or veterinary criteria suitable for the condition being treated or prevented.
- Suitable dosages for Bi modulators are generally as described above. Dosages and methods of administration of any additional agent(s) (e.g., anti-inflammatory and/or analgesic agents) can be found, for example, in the manufacturer's instructions or in the Physician's Desk Reference. In certain embodiments, combination administration results in a reduction of the dosage of the additional agent required to produce a therapeutic effect (i.e., a decrease in the minimum therapeutically effective amount). Thus, preferably, the dosage of additional agent in a combination or combination treatment method of the invention is less than the maximum dose advised by the manufacturer for administration of the agent without combination with a compound of Formula I.
- any additional agent(s) e.g., anti-inflammatory and/or analgesic agents
- combination administration results in a reduction of the dosage of the additional agent required to produce a therapeutic effect (i.e., a decrease in the minimum therapeutically effective amount).
- the dosage of additional agent in a combination or combination treatment method of the invention is less than the maximum dose advised by the manufacturer for administration of the
- this dose is less than 3 ⁇ , even more preferably less than Vi, and highly preferably less than ! ⁇ of the maximum dose, while most preferably the dose is less than 10% of the maximum dose advised by the manufacturer for administration of the agent(s) when administered without combination administration as described herein.
- the dose of compound as provided herein needed to achieve the desired effect may similarly be affected by the dose and potency of the additional agent.
- the present invention provides a variety of non-pharmaceuticai in vitro and in vivo uses for the compounds provided herein.
- such compounds may be labeled and used as probes for the detection and localization of B, (in samples such as cell preparations or tissue sections, preparations or fractions thereof).
- compounds provided herein that comprise a suitable reactive group may be used in photoaffinity labeling studies of receptor binding sites.
- compounds provided herein may be used as positive controls in assays for receptor activity, as standards for determining the ability of a candidate agent to bind to B i, or as radiotracers for positron emission tomography (PET) imaging or for single photon emission computerized tomography (SPECT).
- PET positron emission tomography
- SPECT single photon emission computerized tomography
- a compound may be labeled using any of a variety of well known techniques (e.g., radiolabeled with a radionuclide such as tritium, as described herein), and incubated with a sample for a suitable incubation time (e.g., determined by first assaying a time course of binding). Following incubation, unbound compound is removed (e.g., by washing), and bound compound detected using any method suitable for the label employed (e.g., autoradiography or scintillation counting for radiolabeled compounds; spectroscopic methods may be used to detect luminescent groups and fluorescent groups).
- a radionuclide such as tritium, as described herein
- a matched sample containing labeled compound and a greater (e.g., 10-foid greater) amount of unlabeled compound may be processed in the same manner.
- a greater amount of detectable label remaining in the test sample than in the control indicates the presence of Bj in the sample.
- Detection assays including receptor autoradiography (receptor mapping) of Bi in cultured cells or tissue samples may be performed as described by Kuhar in sections 8.1.1 to 8.1.9 of Current Protocols in Pharmacology (1998) John Wiley & Sons, New York. The following Examples are offered by way of illustration and not by way of limitation.
- Mass spectroscopy data in the following Examples is Electrospray MS, obtained in positive ion mode using a Micromass Time-of-Flight LCT (Micromass, Beverly MA), equipped with a Waters 600 pump (Waters Corp.; Milford, MA), Waters 996 photodiode array detector, and a Gilson 215 autosampler (Gilson, Inc.; Middleton, WI). MassLynx (Advanced Chemistry Development, Inc; Toronto, Canada) version 4.0 software with OpenLynx Global ServerTM, OpenLynxTM and AutoLynxTM processing is used for data collection and analysis.
- Sample volume of 1 microliter is injected onto a 50x4.6mm Chromolith SpeedROD RP- 18e column (Merck KGaA, Darmstadt, Germany), and eluted using a 2 -phase linear gradient at a flow rate of 6 ml/min. Sample is detected using total absorbance count over the 220-340nm UV range.
- the elution conditions are: Mobile Phase A - 95% water, 5% MeOH with 0.05% TFA; Mobile Phase B - 5% water, 95% MeOH with 0.025% TFA.
- the following gradient is used: 0-0.5 min 10-100%B 3 hold at ] 00%B to ] .2 min, return to I0%B at 1.21 min. inject to inject cycle is 2.15 min.
- Step 4 4-Methoxy-2,6,N-trimethyl-N- ⁇ 2-[2-(9-methyl-3,9-diaza-spiro[5.5]undec-3-yl)'2-oxo- ethoxy] ethyl ⁇ benzenesulfonamide
- a 1.0 M solution of TEA in toluene (0.15 mL) is added to a solution of ⁇ 2-[(4-methoxy-2,6- dimethylbenzenesulfonyl)methylamino]ethoxy ⁇ acetic acid (24 mg, 72 ⁇ mol), 3-methyl-3,9-diaza- spiro[5.5]undecane (10 mg, 60 ⁇ mol), and DMA (0.3 mL) under N 2 .
- a 0.2 M solution of 2-chloro- 1,3-dimethylimidazolidinium chloride in acetonitrile (0.6 mL) is added. The reaction vessel is sealed and the mixture is warmed to 50 0 C for 2.5 h.
- Step I (3S)-Methyl 4-benzyl-6-(iodomethyl)-6-methy!morphofine-3-carboxylate
- step 1 The mixture of diastereomers obtained in step 1 is combined with tributyltin hydride (73 mL, 272 mmol) in 400 mL refhixing toluene.
- AIBN (1 g, 6 mmol) dissolved in 40 mL toluene is added dropwise. After 2 h reflux, the reaction mixture is cooled to it.
- Saturated aqueous KF solution (800 mL) is added and the resulting heterogeneous mixture is filtered through Celite and the so ⁇ d washed with EtOAc.
- the combined solutions are transferred to a separatory funnel and washed twice with 200 mL saturated KF, and then once with 20 mL brine.
- the organic phase is applied to a 10 g silica SPE cartridge and eluted with ⁇ O mL of EtOAc to remove non-polar impurities, followed by 10 mL of 10/1/1 EtOAc/MeOH/TEA to elute the title compound.
- the solvent is removed in vacuo to afford the title compound.
- the IC 50 (determined as described in Example 7) is I micromolar or less.
- RNA is isolated as described by Chomzynski et al. (1987) Anal. Biochem. 762: 156-159.
- a cDNA encoding Bj is cloned from the total RNA by reverse transcriptase-polymerase chain reaction (RT-PCR) with the following oligonucleotides:
- Primer 1 GGCGCTAGCCACCATGGCATCCTGGCCCCCTC (SEQIDNO:!)
- Primer 2 AGCCGTCCCAGATCTGAAC (SEQIDNO:2)
- Primer 3 GATCTGGGACGGCTTGGATG (SEQIDNO:3)
- Primer 4 CGGAGCTCTTAATTCCGCCAGAAAAGTTGGA (SEQ ⁇ DNO:4)
- Primer pairs 1 & 2 and 3 & 4 are used to generate overlapping cDNA fragments corresponding to the entire protein coding sequence of cynomolgus macaque Bj cDNA are isolated and linked to form the full-length coding sequence (GenBank Accession Number AY788905), The construct is cloned into pcDNA 3.1 (Invitrogen, Carlsbad, CA) and transfected into Chinese hamster ovary (CHO) ceils using Lipofectamine (Invitrogen), resulting in cynomolgus macaque B r expressing CHO cells.
- the construct is cloned into pBAKPAK9 (Clontech, Mountain View, CA) and transfected into Sf9 cells to generate clonal baculovirus stocks.
- Clonal cell lines stably expressing the cynomolgus macaque Bi receptor are selected in G418.
- a single clonal line that exhibits high levels of receptor expression is chosen for use in binding and calcium mobilization assays (Examples 6 and 7).
- Clonal baculovirus stocks are used to infect Sf9 cells such that the infected cells express high levels of recombinant Bi receptors. These cells are used in radioligand binding assays (Example 6).
- IMR-90 cells which endogenously express human Bi, are seeded into 24 well plates at 65,000 cells per well, cultured overnight, and then treated for 3 h with 0.2 ng/mL interleukin-1 beta to induce B. expression (Menke, et ai, (1994) J. Biol Chem. 269:21583-86).
- CHO cells stably expressing rat B 5 are seeded into 24 well plates at 200,000 cells per well and cultured overnight. The cells are then washed 3 times with phosphate buffered saline (PBS).
- PBS phosphate buffered saline
- binding buffer 50 mM Tris 7.4, 0.14 mg/mL, bacitracin, and 1 mg/mL BSA
- DMSO concentration 1% by volume; some wells receive DMSO only, and some wells receive DMSO plus 10 ⁇ M desArg 10 KalIidin to define non-specific binding. All wells then receive 0.3 nM (final concentration) [3H]-desArg 10 Kallidin. The plates are allowed to sit for 2 h at room temperature. Cells are then washed three times, and lysed with 400 ⁇ l Ultima Gold scintillation fluid (PerkinElmer; Boston, MA; 20 min incubation). The fluid is then transferred to counting vials counted in a Packard liquid scintillation counter (PerkinElmer).
- the number of counts present in the scintillation fluid is plotted as a function of antagonist compound concentration and fitted to a logistical equation using SigmaPlot (Systat Software, Point Richmond, CA) to determine each compound's IC 50 and K 1 (e.g., as described by Szallasi. et al. (1993) J. Pharmacol Exp. Ther. 25(5:678-83).
- Sf9 cells infected with a baculovirus carrying the coding sequence for cynomolgus macaque Bi are harvested by centrifugation and frozen at -SO 0 C. Pellets are subsequently resuspended on ice in Tris buffered saline (TBS; 50 mM Tris (pH 7.4), 120 mM NaCl), and cells are homogenized using a polytron for 30 seconds. The crude membrane fraction is collected by centrifugation at 20,000 rpm. Membranes are washed two times with TBS and collected by centrifugation each time.
- TBS Tris buffered saline
- Protein content of the membranes is determined after the last wash and the concentration is adjusted to 0.7 ⁇ g/uL with binding buffer (50 mM Tris 7.4, 0.14 mg/mL bacitracin, and 1.0 mg/mL BSA).
- binding buffer 50 mM Tris 7.4, 0.14 mg/mL bacitracin, and 1.0 mg/mL BSA.
- Some wells receive DMSO only, and some wells receive DMSO plus 10 ⁇ M desArg 10 Kallidin to define non-specific binding.
- the 96 well plates are allowed to sit for 2 h at room temperature.
- Membrane proteins are then harvested by filtration onto GF/C filtermats (PerkinElmer) pre-soaked for 1 hr in 0.5 % polyethylenimine. After filtration, filters are dried and then counted in a Beta plate counter. The number of counts present in the scintillation fluid is plotted as a function of antagonist compound concentration and Fitted to a logistical equation using SigmaPlot (Systat Software, Point Richmond. CA) to determine each compound's IC 5 0 and K 1 (e.g., as described by Szallasi, et al. (1993) J. Pharmacol. Exp. Th ⁇ r. 266:678-83).
- EXAMPLE 7 CALCIUM MOBILIZATION ASSAY This Example illustrates representative calcium mobilization assays for use in evaluating test compounds for agonist and antagonist activity.
- Cynomolgus macaque B r expressing CHO cells (Example 5) are plated in a 96 well plate. The cells are cultured for 1 day, after which culture media is emptied from the plate and replaced with 50 ⁇ l of KRH (Krebs-Ringer HEPES buffer: 25 mM HEPES, 5 mM KCl 3 0.96 mM NaH 2 PO 4 , 1 mM MgSO 4 , 2 mM CaCl 2 , 5 mM glucose, 1 rnM probenecid, pH 7.4) supplemented with the calcium- sensitive fluorescent dye Fluo4-AM (5 ⁇ g/ml; Teflabs, Austin, TX). The cells are then incubated at 37 0 C in an environment containing 5% CO 2 . After the 1 hour incubation, the dye solution is removed from the plate, the plate is washed once with KRH, and 100 ⁇ L KRH is added.
- KRH Krebs-Ringer HEP
- y is the maximum fluorescence signal
- x is the concentration of the B
- agonist is the E max
- b corresponds to the EC 50 value
- c is the Hill coefficient.
- this assay is also used to assess Bi agonist activity of the test compound. DETERMINATION OF ANTAGONIST ACTIVITY
- test compounds are added to the cell plate prepared as described above in 100 ⁇ L KRH + 2% DMSO, such that the final volume in each well is 200 microliters and the final DMSO concentration is 1%.
- the EC 50 concentration of desArg 10 Kaliidin is then added to each well of plates containing test compound to determine the extent to which each test compound inhibits an agonist-induced Bi response.
- the maximum fluorescent response is plotted as a function of test compound concentration in order to determine the IC 50 (concentration required to inhibit 50% of the effect of agonist) for each compound at B] .
- Antagonists of B decrease this response by at least about 20%, preferably by at least about 50%, and most preferably by at least 80%, as compared to matched control (i.e., cells treated with desArg ! °Kallidin at the EC 50 concentration in the absence of test compound), at a concentration of I O micromoiar or less, preferably 1 micromolar or less.
- the data is analyzed as follows. First, the average maximum relative fluorescent unit (RFU) response from negative control wells (no agonist) is subtracted from the maximum response detected for each of the other experimental wells. Second, average maximum RFU response is calculated for the positive control wells (agonist wells). Then, percent inhibition for each compound tested is calculated using the equation:
- the % inhibition data is plotted as a function of test compound concentration and test compound IC 5O is determined using a linear regression in which x is ln(concentration of test compound) and y is ln( ⁇ ercent inhibition/( 100 - percent inhibition). Data with a percent inhibition that is greater than 90% or less than 15% are rejected and are not used in the regression.
- the IC 50 is
- This Example illustrates the evaluation of compound toxicity using a Madin Darby canine kidney (MDCK) cell cytotoxicity assay. 1 ⁇ L of test compound is added to each well of a clear bottom 96-wel! plate (Packard,
- MDCK cells ATCC no. CCL-34 (American Type Culture Collection, Manassas, VA), are maintained in sterile conditions following the instructions in the ATCC production information sheet. Confluent MDCK cells are trypsinized, harvested, and diluted to a concentration of 0.1 x 10 6 cells/mL with warm (37°C) medium (VITACELL Minimum Essential Medium Eagle, ATCC catalog # 30-
- ATP-LITE-M Luminescent ATP detection kit is generally used according to the manufacturer's instructions to measure ATP production in treated and untreated MDCK cells.
- PACKARD ATP LITE-M reagents are allowed to equilibrate to room temperature. Once equilibrated, the lyophilized substrate solution is reconstituted in 5.5 mL of substrate buffer solution (from kit). Lyophilized ATP standard solution is reconstituted in deionized water to give a 10 mM stock.
- 10 ⁇ L of serially diluted PACKARD standard is added to each of the standard curve control wells to yield a final concentration in each subsequent well of 200 nM, 100 nM, 50 nM, 25 nM, and 12.5 nM.
- PACKARD substrate solution 50 ⁇ L is added to all wells, which are then covered, and the plates are shaken at approximately 700 rpm on a suitable shaker for 2 min.
- a white PACKARD sticker is attached to the bottom of each plate and samples are dark adapted by wrapping piates in foil and placing in the dark for 10 min.
- Luminescence is then measured at 22°C using a luminescence counter (e.g., PACKARD TOPCOUNT Microplate Scintillation and Luminescence Counter or TECAN SPECTRAFLUOR PLUS), and ATP levels calculated from the standard curve. ATP levels in cells treated with test compound(s) are compared to the levels determined for untreated cells.
- a luminescence counter e.g., PACKARD TOPCOUNT Microplate Scintillation and Luminescence Counter or TECAN SPECTRAFLUOR PLUS
- Cells treated with 10 ⁇ M of a preferred test compound exhibit ATP levels that are at least 80%, preferably at least 90%, of the untreated cells.
- ATP levels that are at least 80%, preferably at least 90%, of the untreated cells.
- cells treated with preferred test compounds exhibit ATP levels mat are at least 50%, preferably at least 80%, of the ATP levels detected in untreated cells.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
Abstract
N-oxide aryl sulfones and sulfoxides of Formula (I) are provided: in which the variables are as described herein. Such compounds may be used to modulate bradykinin receptor activity in vivo or in vitro, and are particularly useful in the treatment of conditions responsive to Bi modulation in humans, domesticated companion animals and livestock animals, including inflammation and pain. Pharmaceutical compositions and methods for using them to treat such disorders are provided, as are methods for using such ligands for receptor localization studies and various in vitro assays.
Description
N-OXIDE ARYL SULFONES AND SULFOXIDES
FIELD OF THE INVENTION
This invention relates generally to N-oxide aryl sulfones and sulfoxides, and to the use of such compounds to treat conditions responsive to bradykiπiii receptor-1 (B}) modulation. The invention further relates to the use of such compounds as reagents for the identification of other agents that bind to B5, and as probes for the detection and localization of B].
CROSS-REFERENCE TO RELATED APPLICATIONS This application claims priority to U.S. Provisional Application 60/823,265, filed August 23,
2006, which is hereby incorporated by reference in its entirety.
DESCRIPTION OF THE SEQUENCE LISTING
SEQ ID NO: 1 Bj PCR primer number 1
SEQ ID NO:2 Bi PCR primer number 2 SEQ ID NO : 3 B, PCR primer number 3
SEQ ID NO:4 Bj PCR primer number 4
BACKGROUND OF THE INVENTION
Millions of people throughout the world suffer debilitating and incapacitating pain. Current treatments for pain can be beneficial, but such treatments are not completely effective, and typically have undesirable side effects. For example, non-steroidal anti-inflammatory drugs are commonly used, but are only moderately effective against pain, and have serious renal and gastrointestinal side effects at high doses. Opiates, such as morphine, are potent analgesics, but their usefulness is limited because of adverse side effects, such as physical addictiveness and withdrawal properties, as well as respiratory depression, mood changes, and decreased intestinal motility with concomitant constipation, nausea, vomiting, and alterations in the endocrine and autonomic nervous systems. There is thus a need for effective agents for the treatment of pain.
Bradykinin (BK) is a nonapeptide that functions in cardiovascular homeostasis, contraction and relaxation of smooth muscles, inflammation and pain. The effects of BK are mediated by specific G protein-coupled BK receptors, of which there are at least two distinct subtypes termed Bj and B2. The B2 receptor is expressed constitutively in a variety of tissues. In contrast, the Bj receptor is inducibly expressed in response to pathophysiological conditions such as inflammation, pain, trauma, bacteria! infection, burns and shock.
Accordingly, Bi is a particularly attractive drug target for these and other conditions, and agents that act at this receptor may be targeted specifically to injured tissues, with minimal effects in
normal tissues. In addition, compounds that bind to Bi and/or modulate the activity of Bi also find use as research tools.
There is thus a need in the art for small molecule modulators of B1 activity. The present invention fulfills this need and provides further related advantages.
SUMMARY OF THE INVENTION
The present invention provides N-oxide aryl sulfones and sulfoxides that satisfy Formula I:
Formuia I
 or are a pharmaceutically acceptable salt, solvate (e.g., hydrate) or ester of such a compound.
Within Formula I: A is N, C or CH; = represents a single or double bond;
Ar is phenyl or a 5- or 6-membered heteroaryl. each of which is optionally substituted, and each of which is preferably substituted with from 0 to 4 substituents independently chosen from Rs; Rx and Rγ are: fi) independently chosen from: (a) hydrogen; and
(b) CrC6alkyl, C2-C6alkenyl, C2-C6alkynyl, C,-C6haloalkyl and (CrC8cycloalkyl)Co-C4alkyI, each of which is optionally substituted, and each of which is preferably substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, amino and Ci-Qalkyi; or (ii) taken together to form a 4- to 10-membered carbocycle or a 4- to 10-membered heterocycle, each of which is optionally substituted and each of which is preferably substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, halogen, amino and CrCsaIkyi; Y is a group of the formula (CH2)^Q-(CH2)P, which is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from (i) amino, hydroxy, cyano, Ci-Caalkyl, C2-C6alkenyl, C2-C6alkynyl and Ci-C≤haloalkyl; (ii) substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C^-C6cycloaikyl, and (iii) substituents that are taken together with Rx or Ry to form an optionally substituted 4- to 7- membered carbocycle or 4- to 7-membered heterocycle, wherein:
Q is absent, CH2, CH=CH, O, S or NR10, wherein Ri 0 is hydrogen or d-C6alkyl; and r and p are independently chosen integers ranging from 0 to 6;
W is CrQalkylene that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R2o;
Z is absent, divalent phenyl, NRG or CRERF wherein:
RG is hydrogen, Cj-C6alkyl or taken together with RB or RD to form a 4- to 7-membered heterocycloalkyl that includes W and is optionally substituted; preferably such heterocycloalkyl is substituted with from 0 to 4 substituents independently chosen from R20; RE is hydrogen, Ci-Qalkyl or taken together with R8 to form a 4- to 7-membered heterocycloalkyl that includes W and is optionally substituted; preferably such heterocycloalkyl is substituted with from 0 to 4 substituents independently chosen from R20; and
RF is hydrogen,
 or taken together with RD to form a 4- to 7-membered heterocycloalkyl that includes V and is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R20;
V is absent or Cj-C6a!ky!ene that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from (i) R20 and (ii) groups that are taken together with R0 to form a 4- to 7-membered heterocycle that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R2o;
Each R| is independently chosen from:
(i) halogen, hydroxy, cyano, amino, nitro, aminocarbonyl, aminosulfonyl and -COOH; (ii) C-Qalkyl, C2-C6alkenyl. C2-C6alkynyl, (C3-C8cycloalkyl)Co-C4alkyl, C-Qalkoxy, Cr Qalkylthio, CrC6alkylsuIfinyl, Ci-C6alkoxycarbonyl, C|-C6alkylsulfonylCo-C4alkyI, mono- or di-(C|-C6alkyl)aminoCo-C4alkyl, mono- or di-(Ci-C6alkyl)aminosulfonylCo-C4alkyl, mono- or di-(Ci-C6alkyI)aminocarbonylCo-C4alkyl and (4- to 8-membered heterocycloalkyl)Co- Qalkyl; each of which is optionally substituted and each of which is preferably substituted with from 0 to 6 substituents independently chosen from halogen, hydroxy, cyano and amino; and (iii) groups that are taken together to form a fused carbocyclic ring that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, C;-C6alkyl, Ci-C6haloalkyl, Cj-C6alkoxy, and mono- or di -(Ci -C6alkyl)am inoCo-C4alkyl ;
RB is: (i) hydrogen, C,-C6alkyl, C2-C6alkenyl, C2-C6alkyny!, CrC6haloalkyl, (C3-C3cycloalky])Co-
Cjalkyl, C2-C6alkyi ether, mono- or di-(C|-C6alkyl)aminoCrC4alkyl or (4- to 7-membered heterocycloalkyl)Co-C4alkyL each of which is optionally substituted and each of which is preferably substituted with from 0 to 6 substituents independently chosen from: (a) amino, halogen, hydroxy, cyano and oxo; and (b) C,-C6alkyl, (C3-C8cycloalkyf)C0-C4alkyl, CrC6alkoxy, pheπylC0-C4alkyj and (5- or 6- membered heterocycle)Co-C4aIkyi, each of which is optionally substituted and each of which is preferably substituted with from 0 to 4 substituents independently chosen from
amino, cyano, halogen, hydroxy, Cj-Qalkyl, (C3-C8cycloa!kyl)Co-C4alkyl, Cr C6haloalkyl, CrCόalkoxy, Ci-C6alkoxycarbonyi, mono- or di-(Ci-C6alkyI)ammo, phenyIC0-C4alkyI and phenylCo-Qalkoxy; or
(ii) taken together with RE or R0 to form an optionally substituted heterocycloalkyl; Rc is absent, CrC6alkyl or taken together with RD to from a 4- to 7-menibered heterocycle that is optionally substituted and is preferably substituted with from 0 to 4 substituents independently chosen from R20; RD is CpQalkyl or taken together with Rc, RF, RG or a substituent of V to form an optionally substituted 4- to 7-membered heterocycle; and Each R2O is independently selected from:
(i) hydroxy, oxo, cyano and amino; and
(ii) Ci-Cgalkyl, CrC6alkoxy, mono- or di-(CrC6aikyl)aminoC0-C4alky!, (C3-C 10carbocycle)C0-
Qalkyl and (4- to 10-membered heterocycle)C0-C4aikyl; each of which is optionally substituted, and each of which is preferably substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, Cj-Qaikyi, Q-Qalkoxy, (C3-
Ciocarbocycle)Co-C4alkyl and (4- to 10-membered heterocycle)C0-C4alkyl.
Within certain aspects, N-oxide aryl sulfones and sulfoxides of Formula I, and other Formulas provided herein, are Bt modulators and exhibit a K1 of no greater than 5 micromolar, 2 micrornolar, 1 micromolar, 500 nanomolar, 100 nanomolar, 50 nanomolar or 10 nanomolar in a Bi binding assay and/or have an ECS0 or IC50 value of no greater than 5 micromolar, 2 micromolar, 1 micromolar, 500 nanomolar, 100 nanomolar, 50 nanomolar or 10 nanomolar in an assay for determination of B^ agonist or antagonist activity.
In certain embodiments, B3 modulators provided herein are Bi antagonists; preferably such antagonists exhibit no detectable B] agonist activity, Within certain aspects, N-oxide aryl sulfones and sulfoxides of Formula I are labeled with a detectable marker (e.g., radiolabeled or fluorescein conjugated).
The present invention further provides, within other aspects, pharmaceutical compositions comprising at least one N-oxide aryl sulfone or sulfoxide of Formula I in combination with a physiologically acceptable carrier or excipient. Methods are further provided for inhibiting agonist-induced B1 activity. Within certain such aspects, the inhibition takes place in vitro. Such methods comprise contacting a B| receptor with at least one B] antagonist as described herein, under conditions and in an amount or concentration sufficient to detectably inhibit agonist-induced Bi activity. Within other such aspects, the B5 receptor is in a patient. Such methods comprise contacting cells expressing a B] receptor in a patient with at least one Bi antagonist as described herein in an amount or concentration that would be sufficient to detectably inhibit agonist-induced Bi activity in cells expressing a cloned B] receptor in vitro.
The present invention further provides methods for treating a condition responsive to Bj receptor modulation in a patient, comprising administering to the patient a therapeutically effective amount of at least one N-oxide aiyl sulfone or sulfoxide of Formula I.
Within other aspects, methods are provided for treating pain in a patient, comprising administering to a patient suffering from (or at risk for) pain a therapeutically effective amount of at least one N-oxide aryl sulfone or sulfoxide of Formula ϊ. Pain conditions that may be treated include, but are not limited to, inflammatory pain, acute pain, dental pain, back pain, surgical pain, headache, neuropathic pain, and pain associated with osteoarthritis or trauma.
Within further aspects, the present invention provides methods for determining the presence or absence of Bi in a sample, comprising: (a) contacting a sample with an N-oxide aiyl sulfone or sulfoxide of Formula I under conditions that permit binding of the compound to B,; and (b) detecting a signal indicative of a level of the compound bound to Bi.
In yet another aspect, the present invention provides methods of preparing the compounds disclosed herein, including the intermediates. These and other aspects of the present invention will become apparent upon reference to the following detailed description.
DETAILED DESCRIPTION
As noted above, the present invention provides N-oxide aryl suifones and sulfoxides, which may be used in vitro or in vivo in a variety of contexts, as described herein. TERMINOLOGY
Compounds are generally described herein using standard nomenclature. For compounds having asymmetric centers, it should be understood that (unless otherwise specified) all of the optical isomers and mixtures thereof are encompassed. In addition, compounds with carbon-carbon double bonds may occur in Z- and E- forms, with all isomeric forms of the compounds being included in the present invention unless otherwise specified. If a compound exists in various tautomeric forms, a recited compound is not limited to any one specific tautomer, but rather is intended to encompass all tautomeric forms. Certain compounds are described herein using a general formula that includes variables {e.g., Y, Ar, R)). Unless otherwise specified, each variable within such a formula is defined independently of any other variable, and any variable that occurs more than one time in a formula is defined independently at each occurrence.
The term "N-oxide aryl sulfone or sulfoxide" encompasses all compounds that satisfy Formula I, including compounds in which the Ar moiety is heteroaryl and compounds in which the Ar moiety is aiyl. This term further includes pharmaceutically acceptable salts, solvates (e.g., hydrates) and esters of such compounds. A "pharmaceutically acceptable salt" of a compound recited herein is an acid or base salt that is suitable for use in contact with the tissues of human beings or animals without excessive toxicity or
carcinogenicity, and preferably without irritation, allergic response, or other problem or complication. Such salts include mineral and organic acid salts of basic residues such as amines, as well as alkali or organic salts of acidic residues such as carboxylic acids. Specific pharmaceutically acceptable anions for use in salt formation include, but are not limited to, acetate, 2-acetoxybenzoate, ascorbate, benzoate, bicarbonate, bitartrate, bromide, calcium edetate, carbonate, chloride, citrate, dihydrochloride, diphosphate, edetate, estolate (ethylsuccinate), formate, fumarate, gluceptate, gluconate, glutamate, glycolate, glycollylarsanilate, hexylresorcinate, hydrabamine, hydrobromide, hydrochloride, hydroiodide, hydroxymaleate, hydroxynaphthoate, iodide, isethionate, lactate, lactobionate, malate, maleate, mandelate, methyl bromide, methyl nitrate, methylsulfate, mucate, napsylate, nitrate, pamoate, pantothenate, phenylacetate, phosphate, poϊygalacturonate, propionate, salicylate, stearate, subacetate, succinate, sulfamate, sulfanilate, sulfate, sulfonates including besylate (benzenεsulfoπate), camsylate (cainphorsulfonate), edisylate (ethane-l ,2-disulfonate), esylate (ethanesulfonate), 2-hydroxyethylsulfonate, mesylate (methanesulfonate), triflate (trifluoromethanesulfonate) and tosylate (p-toluenesulfonate), tannate, tartrate, teoclate and triethiodide. Similarly, pharmaceutically acceptable cations for use in salt formation include, but are not limited to ammonium, benzathine, chloroprocaine, choline, diethanolamine, ethyl enediamine, meglumine, procaine, and metals such as aluminum, calcium, lithium, magnesium, potassium, sodium and zinc. Those of ordinary skill in the art will recognize further pharmaceutically acceptable salts for the compounds provided herein. In general, a pharmaceutically acceptable acid or base salt can be synthesized from a parent compound that contains a basic or acidic moiety by any conventional chemical method. Briefly, such salts can be prepared by reacting the free acid or base forms of these compounds with a stoichiometric amount of the appropriate base or acid in water or in an organic solvent, or in a mixture of the two; generally, the use of nonaqueous media, such as ether, ethyl acetate, ethanol, methanol, isopropanol or acetonitrile, is preferred. It will be apparent that each compound provided herein may, but need not, be formulated as a solvate (e.g., a hydrate) or non-covalent complex. In addition, the various crystal forms and polymorphs are within the scope of the present invention. Also provided herein are prodrugs of the compounds provided herein. A "prodrug" is a compound that may not fully satisfy the structural requirements of a formula provided herein, but is modified in vivo, following administration to a patient, to produce a compound within the scope of such formula. For example, a prodrug may be an acylated derivative of a compound as provided herein. Prodrugs include compounds wherein hydroxy, amine or sulfhydryl groups are bonded to any group that, when administered to a mammalian subject, cleaves to form a free hydroxy, amino, or suifhydryl group, respectively. Examples of prodrugs include, but are not limited to, acetate, formate, phosphate and benzoate derivatives of alcohol and amine functional groups within the compounds provided herein. Prodrugs of the compounds provided herein may be prepared by modifying functional groups present in the compounds in such a way that the modifications are cleaved in vivo to yield the parent compounds.
As used herein, the term "alkyl" refers to a straight or branched chain saturated aliphatic hydrocarbon. Alkyl groups include groups having from 1 to 8 carbon atoms (CrCsalkyl), from 1 to 6 carbon atoms (CrC6aiky!) and from 1 to 4 carbon atoms (Ci-C4alkyl)3 such as methyl, ethyl, propyl, isopropyl, n-butyl, sec-butyl, /<?r/-butyi, pentyl, 2-pentyl, isopentyl, neopentyl, hexyi. 2-hexyl, 3-hexyl or 3-metliylpentyl. "C0-C4alkyr' refers to a single covalent bond (C0) or an alkylene group having 1, 2, 3 or 4 carbon atoms; "C0-C2 alky! " refers to a single covalent bond or a methylene or ethylene group.
"Aϋkyieπe" refers to a divalent alkyl group, as defined above. Ci-C4alkylene is an alkylene group having 1, 2, 3 or 4 carbon atoms. "Alkenyl" refers to straight or branched chain alkene groups, which comprise at least one unsaturated carbon-carbon double bond. Alkenyl groups include C2-Qalkenyl, C2-C6alkenyl and C2- C4alkenyl groups, which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively, such as ethenyl, ailyl or isopropenyl. "AlkynyP refers to straight or branched chain alkyne groups, which have one or more unsaturated carbon-carbon bonds, at least one of which is a triple bond. Alkynyl groups include C2-Cgalkynyl, C2-C6aikynyl and C2-C4alkynyl groups, which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively.
A "cycloaikyl" is a saturated or partially saturated cyclic group in which all ring members are carbon, such as cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl and partially saturated variants thereof. Certain cycloaikyl groups are C3-Cscycloalkyl, in which the ring contains from 3 to 8 ring members, all of which are carbon. A "(C3-C3cyc]oalkyI)C0-C4alkyr is a C3-Cgcycloalkyl group linked via a single covalent bond or a CrC4alkyIene group.
By "alkoxy," as used herein, is meant an alkyl group attached via an oxygen bridge (i.e., -O- alkyl). Alkoxy groups include Cj-Qalkoxy and Cj-C4alkoxy groups, which have from 1 to 6 or from
1 to 4 carbon atoms, respectively. Methoxy, ethoxy, propoxy, isopropoxy, n-butoxy, jec-butoxy, terl- butoxy, n-pentoxy, 2-ρentoxy, 3-pentoxy, isopentoxy, neopentoxy, hexoxy, 2-hexoxy, 3-hexoxy, and
3-methyipentoxy are representative alkoxy groups.
Similarly, "alkylthio" refers to an alkyl group as attached via a sulfur bridge (i.e., -S-alkyl). Alkylthio groups include CrC6alkylthio and CrC4alky!thio groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively. "Alkylsulfinyl" refers to groups of the formula -(SO)-alky!, in which the sulfur atom is the point of attachment. Alkylsulfinyl groups include Cj-Cealkylsulfinyl and Ci-C4alkyisulfinyl groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively.
"Alkylsulfonyl" refers to groups of the formula -(SO2)-alkyl, in which the sulfur atom is the point of attachment. Alkylsuifonyl groups include Ci-C6alkylsulfonyl and CrC4alkylsulfonyl groups, which have from 1 to 6 or from 1 to 4 carbon atoms, respectively. The term "Cj-CealkylsuIfonylCo-
C4alkyi refers to a Cj-Cfialkylsulfonyl that is linked via a single covalent bond or a CrC4aIkylene group (i.e., -(C0-C4alky I)-(S O2HC i-C6alky I)).
?
The term "alkanoyP refers to an acyS group (e.g., -(C=O)-alkyl), where attachment is through the carbon of the keto group. Alkanoyl groups inciude Cϊ-Cgalkanoyl, Co-Cgalkanoyl and C2-
C4alkanoyl groups, which have from 2 to 8, 2 to 6 or 2 to 4 carbon atoms, respectively. "Cjalkanoyl" refers to -(C=O)H, which (along with C2-C8alkanoyl) is encompassed by the term "Ci-Qalkanoyl," Ethanoy! is C^alkanoyl.
Similarly, "alkyl ether" refers to a linear or branched ether substituent, Alkyl ether groups include C2-C8alkyl ether, Q-Qalkyl ether and C2-C4a!ky! ether groups, which have 2 to 8. 6 or 4 carbon atoms, respectively. A C2 alkyl ether is -CH2-O-CH3.
The term "alkoxycarbonyl" refers to an alkoxy group linked via a carbonyl (i.e., a group having the general structure -C(=O)-O-alkyl). Alkoxycarbonyl groups include C]-Cg, Ci-C6 and Cr C4alkoxycarbonyl groups, which have from 1 to 8, 6 or 4 carbon atoms, respectively, in the alkyl portion of the group, "Cj alkoxycarbonyl" refers to

"Alkylamino" refers to a secondary or tertiary amine that has the general structure -NH-alkyl or -N(alkyl)(alkyl), wherein each alkyl is selected independently from alkyl, cycloalkyl and (cycloaikyl)alkyl groups. Such groups include, for example, mono- and di-(Ci-C8alkyl)amino groups, in which each Cj-Cgalkyi may be the same or different, as well as mono- and di-(C]-C(;alky1)ammo groups and mono- and di-(CrC4alkyl)amino groups.
"Alkylaminoalkyl " refers to an alkylamino group linked via an alkylene moiety (i.e., a group having the general structure -alkylene— NH-afky! or -aIkylene-N(aikyl)(alkyl)) in which each alkyf is selected independently from alkyl, cycloalkyl and (cycloalkyl)aϊkyl groups. Alkylaminoalkyl groups include, for example, mono- and di-(C]-C3a]kyl)aminoC,-C6alkyl, and mono- and di-(Q- C6alkyl)aminoCi-C4alkyl. "Mono- or di-(Ci-Qalkyl)aminoC0-C4alkyi" refers to a mono- or di-(C,- C6aikyl)amino group linked via a single covalent bond or a C]-C4alkylene group. The following are representative alkylaminoalkyl groups:

It will be apparent that the definition of "alky!" as used in the terms "alkylamino" and "alkylaminoalkyl'1 differs from the definition of "aikyl" used for all other alkyl-containing groups, in the inclusion of cycloalkyl and (cycloalkyl)alkyl groups (e.g., (C3-C7cycloalkyl)C0-C2alkyl).
The term "aminocarbonyl" refers to an amide group (i.e., -C(=O)NH2). "Mono- or di-(Cr C(;alkyl)aminocarbony]Co-C4alkyr refers to an aminocarbonyl group in which one or both hydrogens are replaced with an independently selected Ci-C6alkyl group, and which is linked via a single covalent bond or a CrC4alkylene group.
Similarly, "aminosulfonyl" refers to a sulfonamide group (i.e., -SO2NH2). "Mono- or di-(Cr C6alkyl)aminosulfonylCo-C4aikyl" refers to an aminosulfonyl group in which one or both hydrogens
are replaced with an independently selected CpQalkyl group, and which is linked via a single covalent bond or a Ci-C4alkylene group.
The term "halogen" refers to fluorine, chlorine, bromine or iodine.
The term "oxo," as used herein refers to an oxygen substituent of a carbon atom that results in the formation of a carbonyl group (C=O). An oxo group that is a substituent of a nonaromatic carbon atom results in a conversion of -CH2- to -Cf=O)-. An oxo group that is a substituent of an aromatic carbon atom results in a conversion of -CH- to -C(=O)- and may result in a loss of aromaticity.
A "haloalkyl" is an alkyl group that is substituted with 1 or more independently chosen halogens (e.g., "Ci-Cghaloaikyl" groups have from 1 to 8 carbon atoms; "Q-Qhaloalkyl" groups have from 1 to 6 carbon atoms). Examples of haloalkyl groups include, but are not limited to, mono-, di- or tri-fluoromethyl; mono-, di- or tri-chloromethyi; mono-, di-, tri-, tetra- or penta-fiuoroethyl; mono-, di-, tri-, tεtra- or penta-chloroεthyl; and 1,2,2,2-tetrafluoro-l-trifluoromethyl-ethyI. Typical haloalkyl groups are trifluoromethyl and difluoromethyl.
A dash ("-") that is not between two letters or numbers is used to indicate a point of attachment for a substituent. For example, -C(=O)NH2 is attached through the carbon atom.
A "carbocycle" has from 1 to 3 fused, pendant or spiro rings, each of which has only carbon ring members and each of which may, but need not, be bridged by an alkylene moiety. Typically, a carbocycle that has a single ring contains from 3 to 8 ring members (i.e., C3-Cgcarbocycles); rings having from 4 or 5 to 7 ring members (i.e., C4-C7carbocycles or C5-C7carbocycles) are recited in certain embodiments. Carbocycles comprising fused, pendant or spiro rings typically contain from 9 to 14 ring members. Carbocycles may be optionally substituted with a variety of substituents, as indicated. Unless otherwise specified, a carbocycle may be a cycloalkyl group (i.e., each ring is saturated or partially saturated as described above) or an aryl group (i.e., at least one ring within the group is aromatic). Representative aromatic carbocycles are phenyl, πaphthyi, tetrahydronaphthyl and bi phenyl. In certain embodiments preferred carbocycles have a single ring, such as phenyl and C3- Cscycloalkyl groups.
Certain carbocycles recited herein are
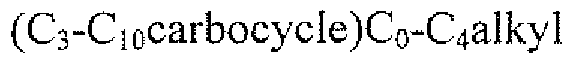 groups (i.e., groups in which a 3- to IO-membered carbocyclic group (which may be cycloalkyl or aryl) is linked via a single covalent bond or a CrC4aIkylene group). Phenyl groups linked via a single covalent bond or Cr C2alkylene group are designated phenyiC0-C4alkyl (e.g., benzyl, l-phenyl-ethyl, 1 -phenyl-propyl and 2-phenyl-ethyl). When substituted, it will be apparent that such groups may be substituted on the ring portion and/or on the alkylene portion of the group. A "phenylC0-C4alkoxy" group is a phenyl ring that is linked via an oxygen atom (i.e., phenoxy) or via a Ci-C4alkoxy group (e.g., benzyloxy).
A "heterocycle" (also referred to herein as a "heterocyclic group") has from 1 to 3 fused, pendant or spiro rings, at least one of which is a heterocyclic ring (i.e., one or more ring atoms is a heteroatom independently chosen from oxygen, sulfur and nitrogen, with the remaining ring atoms being carbon). Typically, a heterocyclic ring comprises 1 , 2, 3 or 4 heteroatoms; within certain
embodiments each heterocyclic ring has 1 or 2 heteroatoms per ring. Each heterocyclic ring generally contains from 3 to 8 ring members (rings having from 4 or 5 to 7 ring members are recited in certain embodiments) and heterocycles comprising fused, pendant or spiro rings typically contain from 9 to 14 ring members. Certain heterocycles comprise a suifur atom as a ring member; in certain embodiments, the sulfur atom is oxidized to SO or SO2. Heterocycles may be optionally substituted with a variety of substituents, as indicated. Certain heterocycles are 4- to 10-membered and comprise one or two rings; in certain embodiments, such heterocycles are monocyclic (e.g., 4- to 8-rnembered, 5- to 8-membered, 4- to 7-membered, or 5- or 6-membered).
Certain heterocycles are heteroaiyl groups (i.e., at least one heterocyclic ring within the group is aromatic), such as a 5- to 10-membered heteroaiyl (which may be monocyclic or bicyclic) or a 5- or 6-membered heteroaryl (e.g., thienyl, imidazolyl, pyridyϊ or pyrimidyl). Other heterocycles are heterocycloalky] groups (i.e., do not comprise an aromatic heterocyclic ring). Certain heterocycles may be linked by a single covalent bond or via an alkylene group, as indicated, for example, by the term "(4- to 8-membered heterocycloalkyl)Co-C4alkyl." A "substituent," as used herein, refers to a molecular moiety that is covalently bonded to an atom within a molecule of interest. For example, a "ring substituent" may be a moiety such as a halogen, alkyl group, haioalkyl group or other group discussed herein that is covalently bonded to an atom (such as a carbon or nitrogen atom) that is a ring member. The term "substitution" refers to replacing a hydrogen atom in a molecular structure with a substituent as described above, such that the valence on the designated atom is not exceeded, and such that a chemically stable compound (i.e., a compound that can be isolated, characterized, and tested for biological activity) results from the substitution.
Groups that are "optionally substituted" are unsubstituted or are substituted by other than hydrogen at one or more available positions, typically 1, 2, 3, 4 or 5 positions, by one or more suitable groups (which may be the same or different). Optional substitution is also indicated by the phrase "substituted with from 0 to X substituents," where X is the maximum number of possible substituents. Certain optionally substituted groups are substituted with from 0 to 2, 3 or 4 independently selected substituents (i.e., are unsubstituted or substituted with up to the recited maximum number of substituents). 11Bi," as used herein, refers to the human Bj bradykinin receptor reported by Menke et al.
(1994) J. Biol. Chem. 2(59:21583-21586, as well as allelic variants thereof and homologues thereof found in other species (e.g., GenBank Accession Number AAX 14712 (Macaca fascicularis)).
The term "Bi agonist" refers to a compound that binds Bi and induces signal transduction mediated by B}. Bi agonists include, for example, bradykinin and kallidin (lysyl-bradykinin), as well as peptide portions or variants of bradykinin or kallidin that bind Bj and retain activity. Representative Bj agonists include, but are not limited to, desArg9bradykinin and desArg'°kallidin.
A "Bi antagonist" is a compound that detectably inhibits signal transduction mediated by B1. Such inhibition may be determined using the representative calcium mobilization assay provided in Example 7. Preferred Bi antagonists have an IC50 Of 5 μM or less in this assay, more preferably 2 μM or less, and still more preferably 1 μM or less, 500 nM or less, 100 nM or less or IO nM or less. In certain embodiments, the B] antagonist is specific for B1 (i.e., the IC50 value in a similar assay performed using the B2 receptor is greater than 2 μM and/or the IC50 ratio (B2/Bi) is at least 10, preferably 100, and more preferably at least 1000). B] antagonists preferably have minimal agonist activity (i.e., induce an increase in the basal activity of Bi that is less than 5% of the increase that would be induced by one EC5O of the peptide agonist desArg'°kallidin, and more preferably have no detectable agonist activity within the assay described in Example 7). B1 antagonists for use as described herein are generally non-toxic. Bi antagonists include neutral antagonists and inverse agonists.
A "neutral antagonist" of Bj is a compound that inhibits the activity of B1 agonist (e.g., desArg'°kallidin) at Bi, but does not significantly change the basal activity of the receptor (i.e., within a calcium mobilization assay as described in Example 7 performed in the absence of agonist, Bj activity is reduced by no more than 10%, more preferably by no more than 5%, and even more preferably by no more than 2%; most preferably, there is no detectable reduction in activity). Neutral antagonists may, but need not, also inhibit the binding of agonist to Bi.
An "inverse agonist" of B1 is a compound that reduces the activity of Bi below its basal activity level in the absence of activating concentrations of agonist. Inverse agonists may also inhibit the activity of agonist at Bi, and/or may inhibit binding of B, agonist to B]. The reduction in basal activity of Bi produced by an inverse agonist may be determined from a calcium mobilization assay. such as the assay of Example 7.
A "therapeutically effective amount" (or dose) is an amount that, upon administration to a patient, results in a discernible patient benefit (e.g., provides detectable relief from a condition being treated). Such relief may be detected using any appropriate criteria. A therapeutically effective amount or dose generally results in a concentration of compound in a body fluid (such as blood. plasma, serum, CSF. synovial fluid, lymph, cellular interstitial fluid, tears or urine) that is sufficient to result in detectable alteration in Br mediated signal transduction (using an assay provided herein). The discernible patient benefit may be apparent after administration of a single dose, or may become apparent following repeated administration of the therapeutically effective dose according to a predetermined regimen, depending upon the indication for which the compound is administered. For the treatment of pain, a discernible patient benefit is generally apparent after administration of a single therapeutically effective dose, although further benefit may become apparent following repeated administrations.
A "patient" is any individual treated with an N-oxide aryl sulfone or sulfoxide as provided herein. Patients include humans, as well as other animals such as companion animals (e.g., dogs and
cats) and livestock. Patients may be experiencing one or more symptoms of a condition responsive to B] modulation or may be free of such symptom(s) {i.e., treatment may be prophylactic in a patient considered to be at risk for the development of such symptoms).
N-OXIDE ARYL SULFONES AND SULFOXIDES
As noted above, the present invention provides N-oxide aryl sulfones and sulfoxides of Formula I that may be used in a variety of contexts, including in the treatment of conditions responsive to B] modulation, as described herein.
Formula I

Such compounds may also be used within in vitro assays (e.g., assays for B1 activity), as probes for detection and localization of Bi and within assays to identify other Bi antagonists. Within certain N-oxide aiyl sulfones and sulfoxides of Formula I, Ar is phenyl, naphthyl, pyridyl or pyrimidinyl, each of which is substituted with from 0 to 4 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, CrC6alkyL CrC6haloalkyl, CpQalkoxy and mono- or
 Within other N-oxide aryl sulfones and sulfoxides of Formula I, Ar is phenyl that is substituted with from 0 to 4 substituents independently chosen from Rj (e.g., substituted with 1, 2 or 3 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, CrQa!kyl, CrC6haloalkyl, C]-C6alkoxy and mono- or di-(Ci-QaIkyl)aminoCo- C4alkyl). Within further N-oxide aryl sulfones and sulfoxides of Formula I5 each substituent (e.g., I5 2 or 3 substituents) represented by Ri is independently chosen from halogen, hydroxy, cyano, amino, nitro, Ci-Cealkyl, CVQ,haloalkyl, CrC6alkoxy, C]-C6haloalkoxy and mono- or di-(Cr C6alkyl)aminoC0-C4alkyl.
Within certain N-oxide aiyl sulfones and sulfoxides, Ar is a group of the formula:

Within the variable designated "Y," the sum of r and p ranges from 1 to 6 for certain N-oxide aryl sulfones and sulfoxides. Within certain such compounds, p is not zero. Certain Y groups satisfy
the formula (C H2X-O-(CH2 )p, which is substituted with from O to 4 substituents independently chosen from amino, hydroxy, cyano, CrC6alkyl, C2-C6alkenyi, C2-C6aikynyl, d-Qhaloalkyl and substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C3-C6cycloalkyl. Representative such Y groups include, for example. -O-CH2-, -CH2-O-CH2-, -O-CH2-CH2-, -CH2- 0-CH2-CH2- and -CH2-CH2-O-CH2-. Other Y groups satisfy the formula -N(R!0)-CH2- Yet another Y group is -CH=CH-.
Within other N-oxide aryl sulfones and sulfoxides of Formula ϊ, Y is a group of the formula (CH2)r-(CH2)p that is substituted with from O to 4 substituents independently chosen from amino, hydroxy, cyano, CrC6alkyl, C2-C6alkenyl, C2-C6alkynyl, Ci-Cghaloaikyl and substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C3-C<;cycloalkyl. Representative such Y groups include, for example, -CH2-, -CH2-CH2-, and -CH2-CH2-CH2-, each of which is optionally substituted as indicated above.
The variable A is N, C, or CH. It will be apparent that if A is N or CH, then == represents a single bond. If =■= represents a double bond, then A is carbon. The variables Rx and Ry of Formula I are, within certain embodiments, independently chosen from hydrogen and CrC6aikyl. Within other embodiments, Rx and Rγ are taken together to form an aromatic group. Representative such aromatic groups include, for example, naphthyl, quinolinyl, phenyl and pyridyl, each of which is optionally substituted as described above. Within further embodiments, Rx and Ry are taken together to form an optionally substituted partially or fully saturated ring. Representative such rings include, for example, piperazine, morpholine, thiomorpholine, piperidine and pyrrolidine.
Certain compounds of Formula I further satisfy Formula II:
Formula II
 wherein Rx and Rγ are:
(i) independently chosen from: (a) hydrogen; and (b) CrC6alkyl, C2-C6alkenyl, C2-C6alkynyl, Ci-Cήhaloalkyl, and (CrCgcycloalkyl)Co-C4alkyl, each of which is substituted with from 0 to
4 substituents independently chosen from oxo, hydroxy, amino and Ci-C4alkyl; or (ii) taken together to form a 4- to I Q-membered heterocycle that is substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, halogen, amino and C3-C8alkyl; and the remaining variables are as described for Formula I. Within certain such compounds, Rx and Ry are taken together to form an optionally substituted 4- to 8-membered heterocycloalkyl. Representative such compounds further satisfy Formula III:
FθrmUlam
 wherein: X is CH2, NR3, O, S, SO or SO2; m and q are independently 0, 1 or 2: R2 represents from 0 to 4 substituents independently chosen from oxo, hydroxy and C|-C6aikyf; R3 is hydrogen, Q-Cβaikyl or CrQaikanoyl; and the remaining variables are as described for Formula L
Certain N-oxide aryl sulfones and sulfoxides of Formula III further satisfy Formula IV or Formula V:
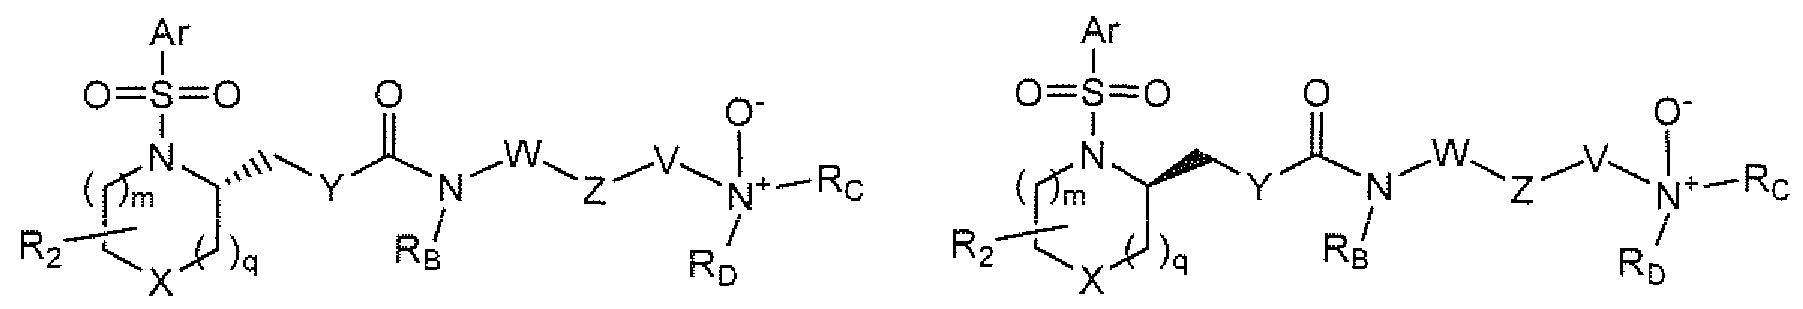
Formula JV Formula V
Within Formulas IH-V, X is CH2 in certain compounds; for other compounds X is O.
Representative
 groups include, for example. ,
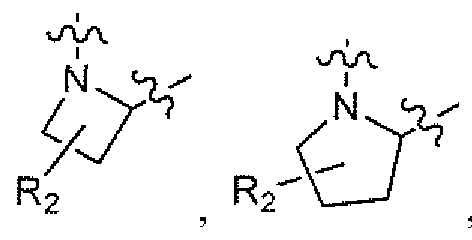
 . Within certain compounds, R2 represents from 0 to 4 substituents independently chosen from C|-C2alkyl (e.g., gem-dimethyl). The variable Rc is absent in certain embodiments. It will be apparent that Rc may only be absent if RD forms a heteroaryl with RF or a substituent of V (and any heteroaryl so formed comprises V and the tetravalent nitrogen atom as ring members).
The variable "Z" is NR0 or CRERp within certain embodiments of Formulas I-V, It will be apparent that RE, RF and RG are intended to be substituents of the base carbon or nitrogen atom (i.e.,
RG RE RF NR0 is "^ ^ and CRERF is "^ ^" ). Within certain such compounds, RE or R0 is taken together with RB to form a 4- to 7-membered heterocycloalkyl that is substituted with from 0 to 4 substituents independently chosen from: (i) hydroxy, oxo, cyano and amino; and (ii) C,-C6alkyl, CrC6alkoxy, mono- or di-(CrC6aIkyl)aminoCo-C4alkyl, (Cj-Ciøcarbocycle)C0-Cιalky] and (4- to 10-membered heteiOcycle)Co-C4alkyl; each of which is substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, CrC6alkyl, C]-C6alkoxy, (C3-C i0carbocycle)C0'C4alkyl and (4- to 10-membered heterocycϊe)Co-Qalkyl. Representative such compounds satisfy Formula VI or Formula VII, in which variables are as described above:

Formula VI Formula VII
Within certain compounds of Formula Vl, s is O, 1 or 2; RD is Ci-Qalkyl; V is absent or Ci- C4alkylene; and the remaining variables are as described above. Within certain compounds of Formula VII, s is O, 1 or 2; RD is CrC6aIkyl; V is C]-C4alkylene; and the remaining variables are as described above. Within certain compounds of Formula VI or Formula VIl, Rx and Rγ are independently hydrogen or Ci-Cealkyl. Other compounds of Formula VI or Formula VlI further satisfy Formula VIII or Formula IX, respectively:

Formula VΪII Formula IX in which m and q are independently O, 1 or 2, such that the sum of m and q is 1, 2 or 3; X is NR3, O, S, SO or SO2; R2 represents from O to 4 substituents independently chosen from oxo, hydroxy and Q- Qalkyl; and the remaining variables are as described for Formula VI. Still further N-oxide aryl sulfones and sulfoxides provided herein satisfy Formula X or
Formula XI:


Within further embodiments of Formulas I- V, the variable "Z" is CRERF, and RF is taken together with RD to form a 4- to 7-membered heterocycloalkyl that is substituted with from 0 to 4 substituents independently chosen from: (i) hydroxy, oxo, cyano and amino; and (ii) CVCgalkyl, C.- Qafkoxy, mono- or di-{C,-C6alkyl)aminoCo-C4alkyi, (C3-Ci 0carbocycle)C0-C4a]ky! and (4- to 10- membered heterocycle)C0-C4alkyI; each of which is substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, C]-C6alkyl, Ci-C6alkoxy, (C3- Ciocarbocycle)Co-C4aikyl and (4- to 10-membered heterocyc!e)Co-C4alkyl. Representative such compounds satisfy Formula XIV:
Formula XIV

Formula XV
 in which m and q are independently 0. 1 or 2, such that the sum of m and q is 1, 2 or 3; X is NR3. O, S, SO or SO2; R2 represents from 0 to 4 substituents independently chosen from oxo, hydroxy and Ci- Cgalkyi; and the remaining variables are as described above.
Within still further embodiments of Formulas I- V, the variable "Z" is NR0 and RQ is hydrogen or CrC6alkyl.
Representative N-oxide aryl sulfones and sulfoxides provided herein include, but are not limited to, those specifically described in the Examples below. It will be apparent that the specific
compounds recited herein are representative only, and are not intended to limit the scope of the present invention. Further, as noted above, all compounds of the present invention may be present as a free acid or base or as a pharmaceutically acceptable salt, hydrate or ester.
As noted above, within certain aspects, compounds provided herein are Bi modulators. In addition, certain compounds provided herein are specific for B1. Bj modulator activity may be confirmed using a calcium mobilization assay, such as the assay described in Example 7, herein.
If desired, binding activity of the compounds provided herein to B] may be confiπned using the representative assay described in Example 6, herein, or using an assay described by Fox et al.
(2005) Br, J. Pharmacol. /44:889-99. Preferred B] modulators exhibit a K1 within such an assay of 5 micromolar or less, more preferably 2 micromolar or less, 1 micromolar or less, 500 nanomolar or less, 100 nanomolar or less or 10 nanomolar or less.
In vivo activity of Bi modulators provided herein may be confirmed using any of a variety of animal models including, but not limited to, those described in the following documents (each of which is hereby incorporated by reference for its disclosure of the recited animal model): Wood et al. (2003) J. Med. Chem. 46: 1803-06 - carrageenan-mduced mechanical pressure hyperalgesia; Cottley et al. (2005) Eur. J. Pharmacol, 527:44-51 - thermal antinociception and carrageenan-induced mechanical pressure hypersensitivity;
Gougat et al. (2004) J. Pharmacol. Exper. Therap. 509:661-669 - UV irradiation-induced thermal hyperalgesia and chronic constriction of the sciatic nerve-induced neuropathy and in ischemia- induced injury; Fox et al. (2005) Br. J. Pharmacol. 744:889-99 - complete Freund's adjuvant-induced mechanical pressure hypersensitivity;
Gabra et ai. (2005) ./ Neuropathol. Exp. Neurol. 64:782-89 - diabetes-induced peripheral neuropathic pain;
Pesquero et al. (2000) Proc. Natl. Acad. Sci. USA 97: 8140-45 - carrageenan-mduced pleurisy
(inflammation around the lung);
Lawson et al. (2005) Eur. J. Pharmacol. 5/4:69-78 - diabetes-induced vascular disease; Hirata et a (2003) Eur. J. Pharmacol. 474:255-60 - ACE inhibitor-induced cough; Mazzuferi et al. (2005) Neuroscience. 135:919-86 - neuronal hyperexcitability in epilepsy.
If desired, compounds provided herein may be evaluated for certain pharmacological properties including, but not limited to, oral bioavailability (preferred compounds are orally bioavailable to an extent allowing for therapeutically effective doses of less than 140 mg/kg, preferably less than 50 mg/kg, more preferably less than 30 mg/kg, even more preferably less than 10 mg/kg, still more preferably less than 1 mg/kg and most preferably less than 0.1 mg/kg), toxicity (a preferred compound is nontoxic when a therapeutically effective amount is administered to a subject), side effects (a preferred compound produces side effects comparable to placebo when a
therapeutically effective amount of the compound is administered to a subject), serum protein binding and in vitro and in vivo half-life (a preferred compound exhibits an in vivo half-life allowing for Q.Ϊ.D. dosing, preferably T.I.D. dosing, more preferably B. LD. dosing, and most preferably once-a- day dosing). In addition, differential penetration of the blood brain barrier may be desirable. Routine assays that are well known in the art may be used to assess these properties, and identify superior compounds for a particular use. For example, assays used to predict bioavailability include transport across human intestinal cell monolayers, including Caco-2 cell monolayers. Penetration of the blood brain barrier of a compound in humans may be predicted from the brain levels of the compound in laboratory animals given the compound {e.g., intravenously). Serum protein binding may be predicted from albumin binding assays. Compound half-life is inversely proportional to the frequency of dosage of a compound. In vitro half-iives of compounds may be predicted from assays of microsomal half-life as described herein.
As noted above, preferred compounds provided herein are nontoxic. In general, the term
"nontoxic" as used herein shall be understood in a relative sense and is intended to refer to any substance that has been approved by the United States Food and Drug Administration ("FDA") for administration to mammals (preferably humans) or, in keeping with established criteria, is susceptible to approval by the FDA for administration to mammals (preferably humans). In addition, a highly preferred nontoxic compound generally satisfies one or more of the following criteria: (1) does not substantially inhibit cellular ATP production; (2) does not significantly prolong heart QT intervals; (3) does not cause substantial liver enlargement, or (4) does not cause substantial release of liver enzymes.
As used herein, a compound that does not substantially inhibit cellular ATP production is a compound that satisfies the criteria set forth in Example 8, herein. In other words, cells treated as described in Example 8 with 100 μM of such a compound exhibit ATP levels that are at least 50% of the ATP levels detected in untreated cells. In more highly preferred embodiments, such cells exhibit
ATP levels that are at least 80% of the ATP levels detected in untreated cells.
A compound that does not significantly prolong heart QT intervals is a compound that does not result in a statistically significant prolongation of heart QT intervals (as determined by electrocardiography) in guinea pigs, minipigs or dogs upon administration of a dose that yields a serum concentration equal to the EC50 or IC50 for the compound. In certain preferred embodiments, a dose of 0.01 , 0.05, 0.1, 0.5, 1 , 5, 10, 40 or 50 mg/kg administered parenterally or orally does not result in a statistically significant prolongation of heart QT intervals. By "statistically significant" is meant results varying from control at the p<0.1 level or more preferably at the p<0.05 level of significance as measured using a standard parametric assay of statistical significance such as a student's T test. A compound does not cause substantial liver enlargement if daily treatment of laboratory rodents (e.g., mice or rats) for 5-10 days with a dose that yields a serum concentration equal to the EC50 or IC50 for the compound results in an increase in liver to body weight ratio that is no more than
100% over matched controls. In more highly preferred embodiments, such doses do not cause liver enlargement of more than 75% or 50% over matched controls. If non-rodent mammals (e.g., dogs) are used, such doses should not result in an increase of liver to body weight ratio of more than 50%, preferably not more than 25%. and more preferably not more than 10% over matched untreated controls. Preferred doses within such assays include 0.01, 0.05. 0.1 , 0.5, 1, 5, 10, 40 or 50 mg/kg administered parenterally or orally.
Similarly, a compound does not promote substantial release of liver enzymes if administration of twice the minimum dose that yields a serum concentration equal to the EC50 or IC5O for the compound does not elevate serum levels of ALT, LDH or AST in laboratory rodents by more than 100% over matched mock-treated controls. In more highly preferred embodiments, such doses do not elevate such serum levels by more than 75% or 50% over matched controls. Alternatively, a compound does not promote substantial release of liver enzymes if, in an in vitro hepatocyte assay, concentrations (in culture media or other such solutions that are contacted and incubated with hepatocytes in vitro) that are equal to the EC50 or IC50 for the compound do not cause detectable release of any of such liver enzymes into culture medium above baseline levels seen in media from matched mock-treated control cells. In more highly preferred embodiments, there is no detectable release of any of such liver enzymes into culture medium above baseline levels when such compound concentrations are five-fold, and preferably ten-fold the EC50 or IC50 for the compound.
In other embodiments, certain preferred compounds do not inhibit or induce microsomal cytochrome P450 enzyme activities, such as CYP 1A2 activity, CYP2A6 activity, CYP2C9 activity, CYP2C19 activity, CYP2D6 activity, CYP2EI activity or CYP3A4 activity at a concentration equal to the EC50 or IC50 for the compound.
Certain preferred compounds are not clastogenic (e.g., as determined using a mouse erythrocyte precursor cell micronucieus assay, an Ames micronucleus assay, a spiral micronucleus assay or the like) at a concentration equal the EC50 or IC50 for the compound. In other embodiments, certain preferred compounds do not induce sister chromatid exchange (e.g., in Chinese hamster ovary cells) at such concentrations.
For detection purposes, as discussed in more detail below, compounds provided herein may be isotopically-Iabeled or radiolabeled. For example, such compounds may have one or more atoms replaced by an atom of the same element having an atomic mass or mass number different from the atomic mass or mass number usually found in nature. Examples of isotopes that can be present in the compounds provided herein include isotopes of hydrogen, carbon, nitrogen, oxygen, phosphorous, fluorine and chlorine, such as 2H, 3H. 11C, 13C. 14C, 15N, 18O, 17O, 31P, 32P, 35S, 18F and 36Cl. in addition, substitution with heavy isotopes such as deuterium (i.e., 2H) can afford certain therapeutic advantages resulting from greater metabolic stability, for example increased in vivo half-life or reduced dosage requirements and, hence, may be preferred in some circumstances.
PREPARATION OF N-OXIDE ARYL SULFONES AND SULFOXIDES
Compounds provided herein may generally be prepared using standard synthetic methods. In general, starting materials are commercially available from suppliers such as Sigma-Aldrich Corp. (St. Louis, MO), or may be synthesized from commercially available precursors using established protocols. By way of example, a synthetic route similar to that shown in any of the following Schemes may be used, together with synthetic methods known in the art of synthetic organic chemistry, or variations thereon appreciated by those skilled in the art. It will be apparent that the reagents and synthetic transformations in the following Schemes and in the art can be readily modified to produce additional compounds of Formula I. Each variable in the following Schemes refers to any group consistent with the description of the compounds provided herein.
When a protecting group is required, an optional deprotection step may be employed. Suitable protecting groups and methodology for protection and deprotection, such as those described in Protecting Groups in Organic Synthesis by T. Greene, are well known. Compounds and intermediates requiring protection/deprotection will be readily apparent.
Certain abbreviations used in the following Schemes and in the Examples include:
AlBN 2,2'-Azobis(2-methylproρionitrile)
DlBAL diisobutylaluminium hydride
DMA N,N-dimethylacetamide
DMAP N,N-dimethyl-4-aminopyiϊdine
DMC 2-chloro-l,3-dimethylimidazolidinium chloride
DMF dimethylformamide
DMSO dimethylsulfoxide
Eq. equivalent(s)
Et ethyl
EtOH ethanol
EtOAc ethyl acetate h hour(s)
1H NMR proton nuclear magnetic resonance
LC-MS liquid chromatography /mass spectrometry mCPBA m-chloroperoxybenzoic acid
MeOH methanol
MHz megahertz
M+l mass + 1 min minute(s)
MS mass spectrometry
NMO N-methylmorpholine N-oxide δ chemical shift
rt room temperature
Ph phenyl
SCX strong cation exchange
TEA triethylamine
TFA trifluoroacetic acid
THF tetrahydrofuran
TPAP tetra-n-propylammonium perruthenate
Scheme 1

Scheme 2

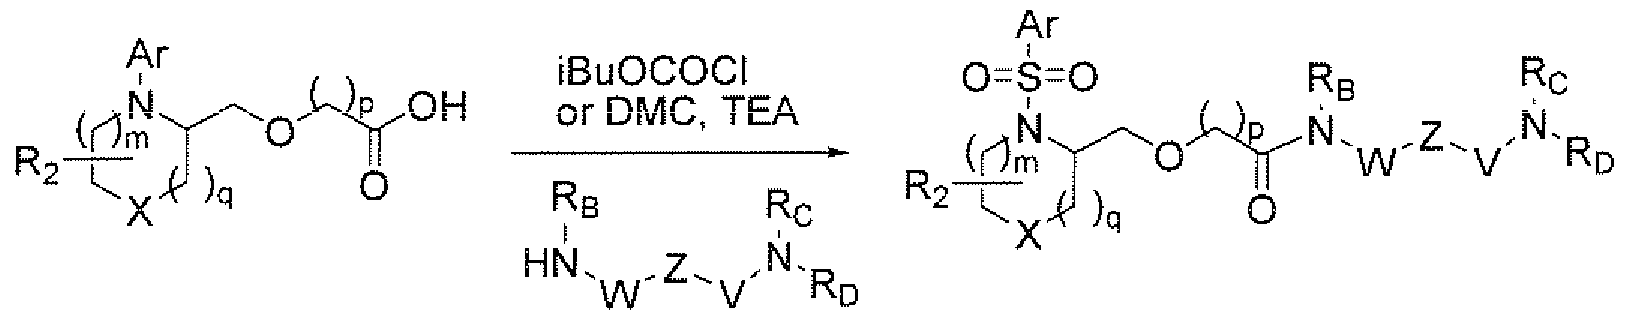
mCPBA

Scheme 3

Scheme 4
 Scheme 5


Scheme 6 RC

mCPBA

Scheme 7
HCi dioxane


In certain embodiments, a compound provided herein may contain one or more asymmetric carbon atoms, so that the compound can exist in different stereo isomeric forms. Such forms can be, for example, racemates or optically active forms. As noted above, all stereoisomers are encompassed by the present invention. Nonetheless, it may be desirable to obtain single enantiomers {i.e., optically active forms). Standard methods for preparing single enantiomers include asymmetric synthesis and resolution of the racemates. Resolution of the racemates can be accomplished, for example, by conventional methods such as crystallization in the presence of a resolving agent, or chromatography using, for example a chiral HPLC column.
Compounds may be radiolabeled by carrying out their synthesis using precursors comprising at least one atom that is a radioisotope. Each radioisotope is preferably carbon (e.g., 14C), hydrogen (e.g., 3H), sulfur (e.g., 3^S) or iodine (e.g., 125I). Tritium labeled compounds may also be prepared catalytically via platinum-catalyzed exchange in tritiated acetic acid, acid-catalyzed exchange in tritiated trifluoroacetic acid, or heterogeneous-catalyzed exchange with tritium gas using che compound as substrate. In addition, certain precursors may be subjected to tritium-halogen exchange with tritium gas, tritium gas reduction of unsaturated bonds, or reduction using sodium borotritide, as appropriate. Preparation of radiolabeled compounds may be conveniently performed by a radioisotope supplier specializing in custom synthesis of radiolabeled probe compounds. PHARMACEUTICAL COMPOSITIONS
The present invention also provides pharmaceutical compositions comprising one or more N- oxide ary! sulfones and sulfoxides provided herein, together with at least one physiologically
acceptable carrier or excipient. Pharmaceutical compositions may comprise, for example, one or more of water, buffers (e.g., neutral buffered saline or phosphate buffered saline), ethanoi, mineral oil, vegetable oil, dimethylsulfoxide, carbohydrates (e.g., glucose, mannose, sucrose or dextrans), rnannitol, proteins, adjuvants, polypeptides or amino acids such as glycine, antioxidants, chelating agents such as EDTA or glutathione and/or preservatives. In addition, other active ingredients may (but need not) be included in the pharmaceutical compositions provided herein.
Pharmaceutical compositions may be formulated for any appropriate manner of administration, including, for example, topical, oral (including, but not limited to, sublingual), nasaϊ, rectal or parenteral administration. The term parenteral as used herein includes subcutaneous, intradermal, intravascular (e.g., intravenous), intramuscular, spina!, intracranial, intrathecal and intraperitoneal injection, as well as any similar injection or infusion technique. In certain embodiments, compositions suitable for oral use are preferred. Such compositions include, for example, tablets, troches, lozenges, aqueous or oily suspensions, dispersible powders or granules, emulsion, hard or soft capsules, or syrups or elixirs. Within yet other embodiments, compositions of the present invention may be formulated as a lyophilizate.
Compositions intended for oral use may further comprise one or more components such as sweetening agents, flavoring agents, coloring agents and/or preserving agents in order to provide appealing and palatable preparations. Tablets contain the active ingredient in admixture with physiologically acceptable excipients that are suitable for the manufacture of tablets. Such excipients include, for example, inert diluents (e.g., calcium carbonate, sodium carbonate, lactose, calcium phosphate or sodium phosphate), granulating and disintegrating agents (e.g., corn starch or alginic acid), binding agents (e.g., starch, gelatin or acacia) and lubricating agents (e.g., magnesium stearate, stearic acid or talc). The tablets may be uncoated or they may be coated by known techniques to delay disintegration and absorption in the gastrointestinal tract and thereby provide a sustained action over a longer period. For example, a time delay material such as glyceryl monosterate or glyceryl distearate may be employed.
Formulations for oral use may also be presented as hard gelatin capsules wherein the active ingredient is mixed with an inert solid diluent (e.g., calcium carbonate, calcium phosphate or kaolin), or as soft gelatin capsules wherein the active ingredient is mixed with water or an oil medium (e.g., peanut oil, liquid paraffin or olive oil).
Aqueous suspensions contain the active material(s) in admixture with excipients suitable for the manufacture of aqueous suspensions. Such excipients include suspending agents (e.g., sodium carboxymethylcellulose, methylceliulose, hydropropylmethylcellulose, sodium alginate, polyvinylpyrrolidone, gum tragacanth and gum acacia); and dispersing or wetting agents (e.g., naturally-occurring phosphatides such as lecithin, condensation products of an alkylene oxide with fatty acids such as polyoxyethylene stearate, condensation products of ethylene oxide with long chain aliphatic alcohols such as heptadecaethyleneoxycetanol, condensation products of ethylene oxide with
partial esters derived from fatty acids and a hexitoi such as polyoxyethylene sorbitol monooleate, or condensation products of ethylene oxide with partial esters derived from fatty acids and hexitoi anhydrides such as polyethylene sorbitan monooleate). Aqueous suspensions may also comprise one or more preservatives, such as ethyl or n-propyl p-hydroxybenzoate, one or more coloring agents, one or more flavoring agents, and one or more sweetening agents, such as sucrose or saccharin.
Oily suspensions may be formulated by suspending the active ingredient(s) in a vegetable oil {e.g., arachis oil, olive oil, sesame oil or coconut oil) or in a mineral oil such as liquid paraffin. The oily suspensions may contain a thickening agent such as beeswax, hard paraffin or cetyl alcohol. Sweetening agents such as those set forth above, and/or flavoring agents may be added to provide palatable oral preparations. Such suspensions may be preserved by the addition of an anti-oxidant such as ascorbic acid.
Dispersible powders and granules suitable for preparation of an aqueous suspension by the addition of water provide the active ingredient in admixture with a dispersing or wetting agent, suspending agent and one or more preservatives. Suitable dispersing or wetting agents and suspending agents are exemplified by those already mentioned above. Additional excipients, such as sweetening, flavoring and coloring agents, may also be present.
Pharmaceutical compositions may also be formulated as oil-in-water emulsions. The oily phase may be a vegetable oil {e.g., olive oil or arachis oil), a mineral oil (e.g., liquid paraffin) or a mixture thereof. Suitable emulsifying agents include naturally-occurring gums (e.g., gum acacia or gum tragacanth), naturally-occurring phosphatides (e.g., soy bean lecithin, and esters or partial esters derived from fatty acids and hexitoi), anhydrides (e.g., sorbitan monoleate) and condensation products of partial esters derived from fatty acids and hexitoi with ethylene oxide (e.g., polyoxyethylene sorbitan monoleate). An emulsion may also comprise one or more sweetening and/or flavoring agents. Syrups and elixirs may be formulated with sweetening agents, such as glycerol, propylene glycol, sorbitol or sucrose. Such formulations may also comprise one or more demulcents, preservatives, flavoring agents and/or coloring agents.
Formulations for topical administration typically comprise a topical vehicle combined with active agent(s), with or without additional optional components. Suitable topical vehicles and additional components are well known in the art, and it will be apparent that the choice of a vehicle will depend on the particular physical form and mode of delivery. Topical vehicles include water; organic solvents such as alcohols (e.g., ethanol or isopropyl alcohol) or glycerin; glycols (e.g., butylene, isoprene or propylene glycol); aliphatic alcohols (e.g., lanolin); mixtures of water and organic solvents and mixtures of organic solvents such as alcohol and glycerin; lipid-based materials such as fatty acids, acylglycerols (including oils, such as mineral oil, and fats of natural or synthetic origin), phosphoglycerides, sphingolipids and waxes; protein-based materials such as collagen and gelatin; silicone-based materials (both non-volatile and volatile): and hydrocarbon-based materials
such as mϊcrosponges and polymer matrices. A composition may further include one or more components adapted to improve the stability or effectiveness of the applied formulation, such as stabilizing agents, suspending agents, emulsifying agents, viscosity adjusters, gelling agents, preservatives, antioxidants, skin penetration enhancers, moisturizers and sustained release materials. Examples of such components are described in Martindaie— The Extra Pharmacopoeia (Pharmaceutical Press, London 1993) and Martin (ed.), Remington's Pharmaceutical Sciences. Formulations may comprise microcapsules, such as hydroxymethylcellulose or gelatin-microcapsules, liposomes, albumin microspheres, microemuisions, nanoparticles or nanocapsules.
A topical formulation may be prepared in a variety of physical forms including, for example, sofids, pastes, creams, foams, lotions, gels, powders, aqueous liquids and emulsions. Typical modes of delivery for topical compositions include application using the fingers; application using a physical applicator such as a cloth, tissue, swab, stick or brush; spraying (including mist, aerosol or foam spraying); dropper application; sprinkling; soaking; and rinsing. Controlled release vehicles can also be used. A pharmaceutical composition may be prepared as a sterile injectible aqueous or oleaginous suspension. The compound(s) provided herein, depending on the vehicle and concentration used, can either be suspended or dissolved in the vehicle. Such a composition may be formulated according to the known art using suitable dispersing, wetting and/or suspending agents such as those mentioned above. Among the acceptable vehicles and solvents that may be employed are water, 1,3-butanedioi, Ringer's solution and isotonic sodium chloride solution. In addition, sterile, fixed oils may be employed as a solvent or suspending medium. For this purpose any bland fixed oil may be employed, including synthetic mono- or diglycerides. In addition, fatty acids such as oleic acid find use in the preparation of injectible compositions, and adjuvants such as local anesthetics, preservatives and/or buffering agents can be dissolved in the vehicle. Compounds may also be formulated as suppositories (e g., for rectal administration). Such compositions can be prepared by mixing the drug with a suitable non-irritating excipient that is solid at ordinary temperatures but liquid at the rectal temperature and will therefore melt in the rectum to release the drug. Suitable excipients include, for example, cocoa butter and polyethylene glycols.
Pharmaceutical compositions may be formulated for release at a pre-determined rate. Instantaneous release may be achieved, for example, via sublingual administration (i.e., administration by mouth in such a way that the active ingredient(s) are rapidly absorbed via the blood vessels under the tongue rather than via the digestive tract). Controlled release formulations (i.e., formulations such as a capsule, tablet or coated tablet that slows and/or delays release of active ingredient(s) foliowing administration) may be administered by, for example, oral, rectal or subcutaneous implantation, or by implantation at a target site. In general, a controlled release formulation comprises a matrix and/or coating that delays disintegration and absorption in the gastrointestinal tract (or implantation site) and thereby provides a delayed action or a sustained action
over a longer period. One type of control led-release formulation is a sustained-release formulation, in which at least one active ingredient is continuously released over a period of time at a constant rate. Preferably, the therapeutic agent is released at such a rate that blood (e.g., plasma) concentrations are maintained within the therapeutic range, but below toxic levels, over a period of time that is at least 4 hours, preferably at least 8 hours, and more preferably at least 12 hours. Such formulations may generally be prepared using well known technology and administered by, for example, oral, rectal or subcutaneous implantation, or by implantation at the desired target site. Carriers for use within such formulations are biocompatible, and may also be biodegradable; preferably the formulation provides a relatively constant level of modulator release. The amount of modulator contained within a sustained release formulation depends upon, for example, the site of implantation, the rate and expected duration of release and the nature of the condition to be treated or prevented.
Controlled release may be achieved by combining the active ingredient(s) with a matrix material that itself alters release rate and/or througii the use of a controlled-release coating. The release rate can be varied using methods well known in the art, including (a) varying the thickness or composition of coating, (b) altering the amount or manner of addition of plasticizer in a coating, (c) including additional ingredients, such as release-modifying agents, (d) altering the composition, particle size or particle shape of the matrix, and (e) providing one or more passageways through the coating. The amount of modulator contained within a sustained release formulation depends upon, for example, the method of administration (e.g., the site of implantation), the rate and expected duration of release and the nature of the condition to be treated or prevented.
The matrix material, which itself may or may not serve a controlled-release function, is generally any material that supports the active ingredient(s). For example, a time delay material such as glyceryl monosterate or glyceryl distearate may be employed. Active ingredient(s) may be combined with matrix material prior to formation of the dosage form (e.g., a tablet). Alternatively, or in addition, active ingredient(s) may be coated on the surface of a particle, granule, sphere, microsphere, bead or pellet that comprises the matrix material. Such coating may be achieved by conventional means, such as by dissolving the active ingredient(s) in water or other suitable solvent and spraying. Optionally, additional ingredients are added prior to coating (e.g., to assist binding of the active ingredient(s) to the matrix material or to color the solution). The matrix may then be coated with a barrier agent prior to application of controlled-release coating. Multiple coated matrix units may, if desired, be encapsulated to generate the final dosage foπn.
In certain embodiments, a controlled release is achieved through the use of a controlled release coating (i.e., a coating that permits release of active ingredient(s) at a controlled rate in aqueous medium). The controlled release coating should be a strong, continuous film that is smooth, capable of supporting pigments and other additives, non-toxic, inert and tack-free. Coatings that regulate release of the modulator include pH-independent coatings, pH-dependent coatings (which may be used to release modulator in the stomach) and enteric coatings (which allow the formulation
to pass intact through the stomach and into the small intestine, where the coating dissolves and the contents are absorbed by the body). It will be apparent that multiple coatings may be employed (e.g., to allow release of a portion of the dose in the stomach and a portion further along the gastrointestinal tract). For example, a portion of active ingredient(s) may be coated over an enteric coating, and thereby released in the stomach, while the remainder of active ingredient(s) in the matrix core is protected by the enteric coating and released further down the GI tract. pH dependent coatings include, for example, shellac, cellulose acetate phthalate, polyvinyl acetate phthalate, hydroxypropylmethylcellulose phthalate, methacrylic acid ester copolymers and zein.
In certain embodiments, the coating is a hydrophobic material, preferably used in an amount effective to slow the hydration of the gelling agent following administration. Suitable hydrophobic materials include alkyl celluloses (e.g., ethylcellulose or carboxymethylcellulose), cellulose ethers, cellulose esters, acrylic polymers (e.g., po!y(acrylic acid), poiy(methacrylic acid), acrylic acid and methacrylic acid copolymers, methyl methacrylate copolymers, ethoxy ethyl methacrylates, cyanoethyl methacrylate, methacrylic acid alkamide copolymer, poly(methyl methacrylate), polyacrylamide, ammonio methacrylate copolymers, aminoalkyi methacrylate copolymer, ρoly(methacrylic acid anhydride) and glycidyl methacrylate copolymers) and mixtures of the foregoing. Representative aqueous dispersions of ethylceilulose include, for example,
AQUACOAT® (FMC Corp., Philadelphia, PA) and SURELEASE® (Colorcon, Inc., West Point,
PA), both of which can be applied to the substrate according to the manufacturer's instructions. Representative acrylic polymers include, for example, the various EUDRAGIT® (Rohm America,
Piscataway, NJ) polymers, which may be used singly or in combination depending on the desired release profile, according to the manufacturer's instructions.
The physical properties of coatings that comprise an aqueous dispersion of a hydrophobic material may be improved by the addition or one or more plasticizers. Suitable plasticizers for alkyl celluloses include, for example, dibutyi sebacate, diethyl phthalate, triethyl citrate, tributyl citrate and triacetin. Suitable plasticizers for acrylic polymers include, for example, citric acid esters such as triethyl citrate and tributyl citrate, dibutyi phthalate, polyethylene glycols, propylene glycol, diethyl phthalate, castor oii and triacetin.
Controlled-release coatings are generally applied using conventional techniques, such as by spraying in the form of an aqueous dispersion. If desired, the coating may comprise pores or channels to facilitate release of active ingredient. Pores and channels may be generated by weil known methods, including the addition of organic or inorganic material that is dissolved, extracted or leached from the coating in the environment of use. Certain such pore-forming materials include hydrophilic polymers, such as hydroxyalkylcelluloses (e g., hydroxypropylmethylcellulose), cellulose ethers, synthetic water-soluble polymers (e.g., polyvinylpyrrolidone, cross-linked polyvinylpyrrolidone and polyethylene oxide), water-soluble polydextrose, saccharides and polysaccharides and alkali metal salts. Alternatively, or in addition, a controlled release coating may include one or more orifices,
which may be formed my methods such as those described in US Patent Nos. 3,845,770; 4,034,758; 4,077,407; 4,088,864; 4,783,337 and 5,071,607. Controlled-release may also be achieved through the use of tran sdermal patches, using conventional technology {see, e.g., US Patent No. 4,668,232).
Further examples of controlled release formulations, and components thereof, may be found, for example, in US Patent Nos. 4,572,833; 4,587,1 17; 4,606,909; 4,610,870; 4,684,516; 4,777,049;
4,994,276; 4,996,058; 5,128,143; 5,202, 128; 5,376,384; 5,384,133; 5,445,829; 5,510,1 19; 5,618,560;
5,643,604; 5,891,474; 5,958,456; 6,039,980; 6, 143,353; 6,126,969; 6,156,342; 6,197,347; 6,387,394;
6,399,096; 6,437,000; 6,447,796; 6,475,493; 6,491,950; 6,524,615; 6,838,094; 6,905,709; 6,923,984;
6,923,988; and 6,911,217; each of which is hereby incorporated by reference for its teaching of the preparation of controlled release dosage forms.
In addition to or together with the above modes of administration, a compound provided herein may be conveniently added to food or drinking water {e.g., for administration to non-human animals including companion animals (such as dogs and cats) and livestock). Animal feed and drinking water compositions may be formulated so that the animal takes in an appropriate quantity of the composition along with its diet. It may also be convenient to present the composition as a premix for addition to feed or drinking water.
N-oxide aryl sulfones and sulfoxides provided herein are generally administered in a therapeutically effective amount. Preferred systemic doses are no higher than 50 mg per kilogram of body weight per day {e.g., ranging from about 0.001 mg to about 50 mg per kilogram of body weight per day), with oral doses generally being about 5-20 fold higher than intravenous doses {e.g., ranging from 0.01 to 40 mg per kilogram of body weight per day). The amount of active ingredient that may be combined with the carrier materials to produce a single dosage unit will vaiy depending, for example, upon the patient being treated and the particular mode of administration. Dosage units will generally contain from about 10 μg to about 500 mg of an active ingredient. Optimal dosages may be established using routine testing, and procedures that are well known in the art.
Pharmaceutical compositions provided herein may, but need not, further comprise one or more additional pharmaceutical agents, such as an anti-inflammatory agent or analgesic.
Anti-inflammatory agents include, for example, non-steroidal anti-inflammatory drugs
(NSAIDs), non-specific and cyclooxygenase-2 (COX-2) specific cyciooxgenase enzyme inhibitors, gold compounds, corticosteroids, methotrexate, leflunomide, cyclosporine A, IM gold, minocycline, azathioprine, tumor necrosis factor (TNF) receptor antagonists, soluble TNF alpha receptor
(etanercept), anti-TNF alpha antibodies {e.g., infliximab and adaϋmumab), anti-C5 antibodies, interleukin- 1 (IL-I) receptor antagonists {e.g., anakinra or IL-I trap), IL- 18 binding protein, CTLA4-
Ig {e.g., abatacept), anti-human IL-6 receptor monoclonal antibody {e.g., tocilizumab), LFA-3-lg fusion proteins {e.g., alefacept), LFA-I antagonists, anti-VLA4 monoantibody {e.g., natalizumab), anti-CDl la monoclonal antibody, anti-CD20 monoclonal antibody {e.g., rituximab), anti-IL-12 monoclonal antibody, anti-IL-15 monoclonal antibody, CDP 484, CDP 870, chemokine receptor
antagonists, selective iNOS inhibitors, p38 kinase inhibitors, integrin antagonists, angiogenesis inhibitors, and TMI-! dual inhibitors. Further anti-inflammatory agents include meloxicam, rofecoxib, celecoxib, etoricoxib, parecoxib, valciecoxib and tilicoxib.
NSAIDs include, but are not limited to, ibuprofen, flurbiprofen, naproxen or naproxen sodium, diclofenac, combinations of diclofenac sodium and misoprostol, sulindac, oxaprozin, diflunisal, piroxicam, indomethacin, etodolac, fenoprofen calcium, ketoprofen, sodium nabumetone, sulfasalazine, tolmetin sodium, and hydroxychloroquine. One class of NSAIDs consists of compounds that inhibit eye 1 ©oxygenase (COX) enzymes; such compounds include celecoxib and rofecoxib. NSAlDs further include salicylates such as acetylsalicylic acid or aspirin, sodium salicylate, choline and magnesium salicylates, and salsalate, as well as corticosteroids such as cortisone, dexamethasone, methylprednisolone, prednisolone, prednisolone sodium phosphate, and prednisone.
Certain analgesics for use in combination with B] modulators provided herein are also antiinflammatory agents, and are listed above. Other such medications are analgesic agents, including narcotic agents which typically act at one or more opioid receptor subtypes (e.g., μ, K and/or δ), preferably as agonists or partial agonists. Such agents include opiates, opiate derivatives and opioids, as well as pharmaceutically acceptable salts and hydrates thereof. Specific examples of narcotic analgesics include, within preferred embodiments, alfentanil, aϊphaprodine, anileridine, bezitramide, buprenorphine, butorphanol, codeine, diacetyidihydromorphine, diacetylmorphine, dihydrocodeine, diphenoxylate, ethylmorphine, fentanyl, heroin, hydrocodone, hydromorphone, isomethadone, levomethorphan, levorphane, levorphanol, meperidine, metazocine, methadone, methorphan, metopon, morphine, nalbuphine, opium extracts, opium fluid extracts, powdered opium, granulated opium, raw opium, tincture of opium, oxycodone, oxymorphone, paregoric, pentazocine, pethidine, phenazocine, piminodine, propoxyphene, racemethorphan, racemorphan, sulfentanyi, thebaine and pharmaceutically acceptable salts and hydrates of the foregoing agents.
Other examples of narcotic analgesic agents include acetorphine, acetyldihydrocodeine, acetylmethadol, allylprodine, alphracetylmethadol, alphameprodine, alphamethadol, benzethidine, benzyimoφhine, betacetylmethadol, betameprodine, betamethadol, betaprodine, clonitazene, codeine methylbromide, codeine-N-oxide, cyprenoφhine, desomorphine, dextromoramide, diampromide, diethylthiambutene, dihydromorphine, dimenoxadol, dimepheptanol, dimethylthiamubutene, dioxaphetyl buryrate. dipipanone, drotebanol, ethanol, ethylmethylthiambutene, etonitazene, etorphine, etoxeridine, furethidine, hydromoφhinol, hydroxypethidine, ketobemidone, levomoramide, levophenacylmoφhan, methyldesorphine, methyldihydromorphine, moφheridine, moφhine, methylpromide, morphine methylsulfonate, moφhine-N-oxide, myrophin. naloxone, naltyhexone, nicocodeine, πicomoφhine, noracymethadol, norlevoφhanol, normethadone, normoφhine, noφipanone, pentazocaine, phenadoxone, phenampromide, phenomoφhan, phenoperidine,
piritramide, pholcodine, proheptazoine, properidine, prop Iran, racemoramide, thebacon, trimeperidine and the pharmaceutically acceptable salts and hydrates thereof.
Further specific representative analgesic agents include, for example acetaminophen
(paracetamol); aspirin and other NSAlDs described above; NR2B antagonists; capsaicin receptor antagonists; anti-migraine agents; anticonvulsants such as oxcarbazepine and carbamazepine; antidepressants (such as TCAs, SSRIs, SNRIs, substance P antagonists, etc.); spinal blocks; pentazocine/naloxone; meperidine; levorphanol; buprenorphine; hydromorphone; fentanyl; sufentanyi; oxycodone; oxycodone/acetaminophen, nalbuphine and oxymorphone. Still further analgesic agents include CB2-receptor agonists, such as AM 1241, capsaicin receptor antagonists and compounds that bind to the α2δ subunit of voltage-gated calcium channels, such as gabapentin and pregabalin.
Representative anti-migraine agents for use in combination with a Bj modulator provided herein include CGRP antagonists, ergotamines and 5-HT] agonists, such as sumatripan, naratriptan, zolmatriptan and rizatriptan. Pharmaceutical compositions may be packaged for treating conditions responsive to B] modulation {e.g., treatment of pain, inflammation or other disorder(s) recited herein). Packaged pharmaceutical preparations generally comprise a container holding a therapeutically effective amount of a pharmaceutical composition as described above and instructions (e.g., labeling) indicating that the composition is to be used for treating a condition responsive to Bs modulation in a patient (e.g., pain or other disorder as indicated herein). In certain embodiments, a packaged pharmaceutical preparation comprises one or more N-oxide aryl sulfone or sulfoxide provided herein and one or more additional agents in the same package, either in separate containers within the package or in the same container (i.e., as a mixture). Preferred mixtures are formulated for oral administration (e.g., as pills, capsules, tablets or the like). In certain embodiments, the package comprises a label bearing indicia indicating that the components are to be taken together for the treatment of pain.
METHODS OF USE
Within certain aspects, the present invention provides methods for treating a condition responsive to B] modulation in a patient. The patient may be afflicted with such a condition, or may be free of symptoms but considered at risk for developing such a condition. A condition is "responsive to B1 modulation'1 if the condition or symptom(s) thereof are alleviated, attenuated, delayed or otherwise improved by modulation of Bs activity. In general, such methods comprise administering to the patient a therapeutically effective amount of at least one N-oxide aiyl suifone or sulfoxide as provided herein.
Conditions responsive to B1 modulation include, for example pain; inflammation including neuroinflammation (such as atherosclerosis), inflammation associated with airway diseases (e.g., asthma, including allergic asthma, exercise-induced bronchoconstriction, occupational asthma, and other non-allergic asthmas), and inflammatory skin disorders (e.g., psoriasis and eczema)); respiratory
disorders including bronchoconstriction, asthma, chronic obstructive pulmonary disease (e.g., emphysema), chronic cough (including ACE-inhibitor cough), adult respiratory distress syndrome, bronchitis, pneumonia, allergic rhinitis and vasomotor rhinitis; vascular edema (including diabetes- related vascular disease); and epilepsy. Other conditions responsive to Bj modulation include diabetes (e.g., type II or non insulin dependent, as well as diabetic vasculopathy, diabetic neuropathy, diabetic retinopathy, post capillary resistance and symptoms associated with insulitis), seizure disorders (e.g., epilepsy), multiple sclerosis, liver disease, cardiovascular disorders (e.g., atherosclerosis, congestive heart failure and myocardial infarction), neurodegenerative diseases (e.g., Alzheimer's disease and Parkinson's disease) rheumatoid arthritis, infection, cancer, crania! trauma, rhinitis, septic shock, endotoxϊc and pancreatic shock, anaphylaxis, inflammatory bowel disease, irritable bowel syndrome, pancreatitis, cystitis, uveitis, vascular permeability, gingivitis, osteoporosis, benign prostatic hyperplasia, hyperactive bladder, cerebral edema, vasodilation, hypotension associated with sepsis, edema resulting from trauma associated with burns, sprains or fracture, cerebral edema, angiodema, Crohn's disease and ulcerative colitis. Bj modulators may also be used as smooth muscle relaxants for treating spasms of the gastrointestinal tract of uterus. In certain embodiments, the condition responsive to B] modulation is pain or inflammation.
Pain that may be treated using the Bi modulators provided herein may be chronic or acute and includes, but is not limited to, peripheral nerve-mediated pain (especially neuropathic pain, such as pain due to diabetes, postherpetic neuralgia, nerve injury, vulvodynia, root avulsions, painful traumatic neuropathy and painful polyneuropathy). Compounds provided herein may also be used in the treatment of, for example, visceral pain (e.g., pancreatitis, interstitial cystitis and renal colic), persistent hyperalgesia, inflammatory pain, repetitive motion pain, carpel tunnel syndrome, perioperative pain, algesia, oral neuropathic pain, toothache (dental pain), denture pain, ocular pain, postherpetic neuralgia, diabetic neuropathy, chemotherapy- induced neuropathy, reflex sympathetic dystrophy, trigeminal neuralgia, bone and joint pain (e.g., pain associated with osteoarthritis), rheumatoid arthritis, myofascial pain (e.g., muscular injury and fibromyalgia), Guillain-Barre syndrome, meralgia paresthetica, burning-mouth syndrome and/or pain associated with nerve and root damage, including as pain associated with peripheral nerve disorders (e.g., nerve entrapment and brachial plexus avulsions, amputation, peripheral neuropathies including bilateral peripheral neuropathy, tic douloureux, atypical facial pain, nerve root damage, and arachnoiditis). Additional neuropathic pain conditions include causalgia (reflex sympathetic dystrophy - RSD, secondary to injury of a peripheral nerve), neuritis (including, for example, sciatic neuritis, peripheral neuritis, polyneuritis, optic neuritis, postfebrile neuritis, migrating neuritis, segmental neuritis and Gombault's neuritis), neuronitis, neuralgias (e.g., those mentioned above, cervicobrachial neuralgia, cranial neuralgia, geniculate neuralgia, glossopharyngial neuralgia, migranous neuralgia, idiopathic neuralgia, intercostals neuralgia, mammary neuralgia, mandibular joint neuralgia, Morton's neuralgia,
nasociliary neuralgia, occipital neuralgia, red neuralgia, Binder's neuralgia, splenopalatine neuralgia, supraorbital neuralgia and vidian neuralgia), musculoskeletal pain, AIDS-related neuropathy, MS- related neuropathy, central nervous system pain (e.g., pain due to brain stem damage, sciatica, and ankylosing spondylitis), and spinal pain, including spinal cord injury-related pain. Headache, including headaches involving peripheral nerve activity may also be treated as described herein. Such headache pain includes, for example, sinus, cluster (i.e., migranous neuralgia) and tension headaches, migraine, temporomandibular pain and maxillary sinus pain. For example, migraine headaches may be prevented by administration of a compound provided herein as soon as a pre-migrainous aura is experienced by the patient. Further conditions that can be treated as described herein include Charcot's pains, intestinal gas pains, ear pain, heart pain, muscle pain, eye pain, orofacial pain (e.g., odontalgia), abdominal pain, gynaec ©logical pain (e.g., menstrual pain, dysmenorrhoea, pain associated with cystitis, labor pain, chronic pelvic pain, chronic prostitis and endometriosis), acute and chronic back pain (e.g., lower back pain), gout, scar pain, hemorrhoidal pain, dyspeptic pains, pain associated with angina, nerve root pain, "non-painful" neuropathies, complex regional pain syndrome, nomotopic pain and heterotopic pain — including pain associated with carcinoma, often referred to as cancer pain (e.g., in patients with bone cancer), pain (and inflammation) associated with venom exposure (e.g., due to snake bite, spider bite, or insect sting) and trauma associated pain (e.g., post-surgical pain such as postmastectomy, postthoracotomy and stump pain; phantom limb pain, episiotomy pain, pain from cuts, musculoskeletal pain, bruises and broken bones, and burn pain, especially primary hyperalgesia associated therewith). Additional pain conditions that may be treated as described herein include pain associated with respiratory disorders as described above, autoimmune diseases, immunodeficiency disorders, hot flashes, inflammatory bowel disease, gastroesophageal reflux disease (GERD), irritable bowel syndrome and/or inflammatory bowel disease. In certain embodiments, pain treated with B] modulators provided herein is inflammatory pain, acute pain, dental pain, back pain, surgical pain, headache, neuropathic pain or pain from osteoarthritis or trauma.
It will be apparent that compounds provided herein may be administered alone or in combination with one or more additional agents that are suitable for treating the disorder of interest. Within such combination therapy, the compound(s) and additional agent(s) may be present in the same pharmaceutical composition, or may be administered separately in either order. Representative anti-inflammatory agents and analgesics for use in combination therapy include those indicated above.
Within other aspects, B| modulators provided herein may be used within combination therapy for the treatment of conditions involving pain and/or inflammatory components. Such conditions include, for example, autoimmune disorders and pathologic autoimmune responses known to have an inflammatory component including, but not limited to, arthritis (especially rheumatoid arthritis), psoriasis, Crohn's disease, lupus erythematosus, irritable bowel syndrome, tissue graft rejection, and
hyperacute rejection of transplanted organs. Other such conditions include trauma (e.g., injury to the head or spinal cord), cardio- and cerebro-vascular disease and certain infectious diseases. Within such combination therapy, a B] modulator is administered to a patient along with an additional analgesic and/or anti-inflammatory agent. The B; modulator and additional analgesic and/or anti- inflammatory agent may be present in the same phaπnaceutical composition, or may be administered separately in either order.
Administration to the patient can be by way of any means discussed above, including oral, topical, nasal or transdermal administration, or intravenous, intramuscular, subcutaneous, intrathecal, epidural, intracerebroventricular or like injection. Oral administration is preferred in certain embodiments (e.g., formulated as pills, capsules, tablets or the like).
Treatment regimens may vary depending on the compound used and the particular condition to be treated. In genera!, a dosage regimen of 4 times daily or less is preferred, with 1 or 2 times daily particularly preferred. It will be understood, however, that the specific dose and treatment regimen for any particular patient will depend upon a variety of factors including the activity of the specific compound employed, the age, body weight, general health, sex and diet of the patient, the time of administration, the route of administration, the rate of excretion, any drug combination and the severity of the particular disease undergoing therapy. Dosages are generally as described above; in general, the use of the minimum dose sufficient to provide effective therapy is preferred. Patients may generally be monitored for therapeutic effectiveness using medical or veterinary criteria suitable for the condition being treated or prevented.
Suitable dosages for Bi modulators (either alone or within such combination therapy) are generally as described above. Dosages and methods of administration of any additional agent(s) (e.g., anti-inflammatory and/or analgesic agents) can be found, for example, in the manufacturer's instructions or in the Physician's Desk Reference. In certain embodiments, combination administration results in a reduction of the dosage of the additional agent required to produce a therapeutic effect (i.e., a decrease in the minimum therapeutically effective amount). Thus, preferably, the dosage of additional agent in a combination or combination treatment method of the invention is less than the maximum dose advised by the manufacturer for administration of the agent without combination with a compound of Formula I. More preferably this dose is less than 3Λ, even more preferably less than Vi, and highly preferably less than !Λ of the maximum dose, while most preferably the dose is less than 10% of the maximum dose advised by the manufacturer for administration of the agent(s) when administered without combination administration as described herein. It will be apparent that the dose of compound as provided herein needed to achieve the desired effect may similarly be affected by the dose and potency of the additional agent. Within separate aspects, the present invention provides a variety of non-pharmaceuticai in vitro and in vivo uses for the compounds provided herein. For example, such compounds may be labeled and used as probes for the detection and localization of B, (in samples such as cell
preparations or tissue sections, preparations or fractions thereof). Tn addition, compounds provided herein that comprise a suitable reactive group (such as an aryl carbonyl, nitro or azide group) may be used in photoaffinity labeling studies of receptor binding sites. In addition, compounds provided herein may be used as positive controls in assays for receptor activity, as standards for determining the ability of a candidate agent to bind to Bi, or as radiotracers for positron emission tomography (PET) imaging or for single photon emission computerized tomography (SPECT). Such methods can be used to characterize Bi receptors in living subjects. For example, a compound may be labeled using any of a variety of well known techniques (e.g., radiolabeled with a radionuclide such as tritium, as described herein), and incubated with a sample for a suitable incubation time (e.g., determined by first assaying a time course of binding). Following incubation, unbound compound is removed (e.g., by washing), and bound compound detected using any method suitable for the label employed (e.g., autoradiography or scintillation counting for radiolabeled compounds; spectroscopic methods may be used to detect luminescent groups and fluorescent groups). As a control, a matched sample containing labeled compound and a greater (e.g., 10-foid greater) amount of unlabeled compound may be processed in the same manner. A greater amount of detectable label remaining in the test sample than in the control indicates the presence of Bj in the sample. Detection assays, including receptor autoradiography (receptor mapping) of Bi in cultured cells or tissue samples may be performed as described by Kuhar in sections 8.1.1 to 8.1.9 of Current Protocols in Pharmacology (1998) John Wiley & Sons, New York. The following Examples are offered by way of illustration and not by way of limitation.
Unless otherwise specified all reagents and solvent are of standard commercial grade and are used without further purification. Using routine modifications, the starting materials may be varied and additional steps employed to produce other compounds provided herein.
EXAMPLES Mass spectroscopy data in the following Examples is Electrospray MS, obtained in positive ion mode using a Micromass Time-of-Flight LCT (Micromass, Beverly MA), equipped with a Waters 600 pump (Waters Corp.; Milford, MA), Waters 996 photodiode array detector, and a Gilson 215 autosampler (Gilson, Inc.; Middleton, WI). MassLynx (Advanced Chemistry Development, Inc; Toronto, Canada) version 4.0 software with OpenLynx Global Server™, OpenLynx™ and AutoLynx™ processing is used for data collection and analysis. MS conditions are as follows: capillary voltage = 3.5 kV; cone voltage = 30 V, desolvation and source temperature = 35O0C and 1200C, respectively; mass range = 181-750 with a scan time of 0.22 seconds and an interscan delay of 0.05 seconds.
Sample volume of 1 microliter is injected onto a 50x4.6mm Chromolith SpeedROD RP- 18e column (Merck KGaA, Darmstadt, Germany), and eluted using a 2 -phase linear gradient at a flow rate of 6 ml/min. Sample is detected using total absorbance count over the 220-340nm UV range. The
elution conditions are: Mobile Phase A - 95% water, 5% MeOH with 0.05% TFA; Mobile Phase B - 5% water, 95% MeOH with 0.025% TFA. The following gradient is used: 0-0.5 min 10-100%B3 hold at ] 00%B to ] .2 min, return to I0%B at 1.21 min. inject to inject cycle is 2.15 min.
EXAMPLE 1. SYNTHESIS OF 4-METHOXY-N,2,6-TR]METHYL-N-{2-[2-(9-METHYL-9-OXIDO-3,9- DIAZASPIRO[S-S]UNDEC-S -YL)-Z-OXOETHOXT]ETHYLIBENZENESULFONAMIDE (COMPOUND 1)

mCPBA

Step 1. N-(2-Hydroxyethyl)-4-methoxy-2,6,N-trimethylbenzenesulfonamide

4-Methoxy-2,6-dimethylbenzenesuIfonyl chloride (12 g, 51 mmol) is added to a solution of 2- (methylamino)ethanol (4.6 mL, 61 mmol), diisopropylethylamine (22 mL, 130 mmol), and CH2CI2 (130 mL) under N2. The reaction vessel is sealed and left to stir for 21 h. The solution is washed with 1 : 1 sat. aq. NH4Cl: H2O (2 x 200 mL) and brine (200 mL). The aqueous layers are extracted with CH2Cl2 (200 mL). The combined organic layers are dried over Na2SO4, filtered, and concentrated to afford the title compound as a pale yellow oil.
Step 2. {2-[(4-Methoxy-2,6-dimethylbenzenesulfonyI)methylamino]ethoxy}acetic acid tert-butyl ester

Sodium hydride, 60% dispersion in mineral oil, (3.7 g, 90 mmol) is added to a solution of (2- hydroxyethyl)-4-methoxy-2,6,N-triraethyIbenzenesulfonamide (14.5 g) and DMF (300 mL) at 0 0C under N2. After 15 min, t-butyl bromoacetate (1 1.5 mL, 78 mmol) is added and the cold bath is removed. After 16 h, the reaction mixture is poured into 50% sat. aqueous NH4CI (400 mL). The solution is extracted with EtOAc (2 x 250 mL). The combined organic layers are washed with 50% sat. aqueous NH4Cl (400 mL), dried over Na2SO4, filtered, and concentrated. Purification by flash column chromatography (5: 1 hexanes: EtOAc, 250 g SiO2) affords the title compound as a pale yellow oil.
Step 3. {2-[(4-Methoxy-2,6-dimethylbenzenesulfonyl)methylamino]ethoxy}acetic acid
OH

{2-[(4-Methoxy-2,6-dimethylbenzenesu]fonyl)methylamino]ethoxy}acetic acid tert-butyl ester (18.9 g, 48.8 mmol) is treated with 4 M HCl in dioxane (60 mL) at rt overnight. The volatiles are removed under reduced pressure to afford a tan solid. 1H NMR (400 MHz, CDCl3) δ: 6.65 (s, 2H), 4.13 (s, 2H), 3.82 (s, 3H), 3.72 (t, 2H), 3.53 (t, 2H), 2.70 (s, 3H), 2.60 (s, 6H).
Step 4. 4-Methoxy-2,6,N-trimethyl-N-{2-[2-(9-methyl-3,9-diaza-spiro[5.5]undec-3-yl)'2-oxo- ethoxy] ethyl } benzenesulfonamide
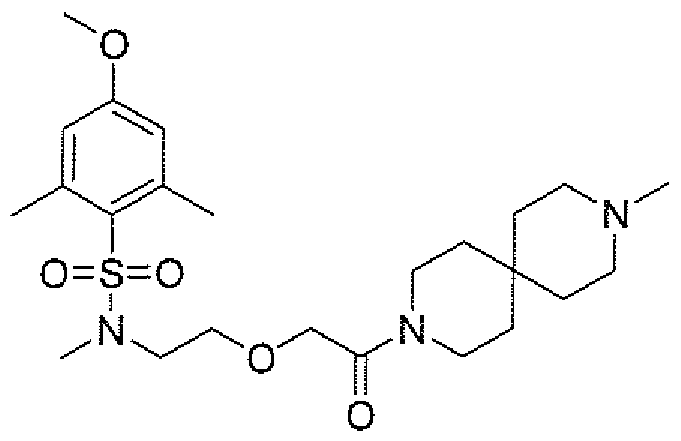
A 1.0 M solution of TEA in toluene (0.15 mL) is added to a solution of {2-[(4-methoxy-2,6- dimethylbenzenesulfonyl)methylamino]ethoxy}acetic acid (24 mg, 72 μmol), 3-methyl-3,9-diaza-
spiro[5.5]undecane (10 mg, 60 μmol), and DMA (0.3 mL) under N2. A 0.2 M solution of 2-chloro- 1,3-dimethylimidazolidinium chloride in acetonitrile (0.6 mL) is added. The reaction vessel is sealed and the mixture is warmed to 50 0C for 2.5 h. After cooling to ambient temperature, 1 M aqueous NaOH (1.5 mL) is added and then the volatiles are removed under reduced pressure. The aqueous residue is extracted with EtOAc (2 x 1.5 mL). The organics are loaded onto two 0.5 g SCX columns. Each column is washed with MeOH (3 x 3 mL) and then eluded with 5:5:2 EtOAc : MeOH : TEA (4.5 mL). Concentration of the organics under reduced pressure afforded the title compound as a pale yellow film. 1H-NMR (400 MHz, CDCl3) & 6.62 (s, 2H), 4.10 (s, 2H), 3.8] (s, 3H), 3.67 (t, 2H), 3.52 (dd, 2H), 3.37 (t, 2H), 3.34 (dd, 2H), 2.78 (s, 3H), 2.60 (s, 6H), 2.35-2.44 (m, 4H), 2.29 (s, 3H), 1.42- 1.60 (m, 8H).
Step 5. 4-Methoxy-236,N-trimethyl-N-{2-[2-(9-methyl-3,9-diaza-spiro[5.5]undec-3-yl)-2-oxo- ethoxyjethyl } benzenesulfonam ide
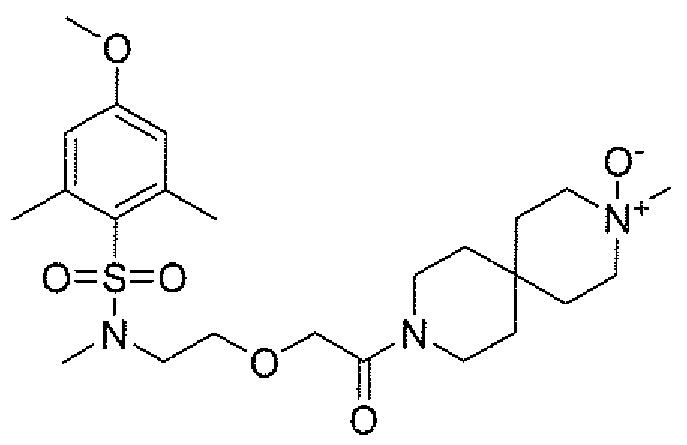
A 0.25 M solution of 3-chloroperbenzoic acid in CH2Cl3 (500 μL, 125 μmol) is added to a 0 °C solution of 4-methoxy-2,6,N-trimethyl-N-{2-[2-(9-methyl-3,9-diaza-spiro[5.5]undec-3-yl)-2-oxo- ethoxy]ethyl}benzenesulfonamide (50 mg, 104 μmol) in CH2Cl2 (500 μL), After 1.5 h, the volatiles are removed under reduced pressure. The resulting residue is purified by thin layer chromatography (eluting with 2% NH4OH in 5 : 1 CH2CI2 : MeOH) to afford the title compound. 1H-NMR (400 MHz, CDCS3) & 6.62 (s, 2H), 4.11 (s, 2H), 3.81 (s, 3H), 3.68 (t, 2H), 3.22-3.60 (m, 12H), 3.28 (s, 3H), 2.77 (s, 3H), 2.59 (s, 6H), 2.16-2.34 (m, 2H), 1.47-1.68 (m, 4H). LC-MS m/∑ (M + H+): 498.3; retention time: 0.9 min. The IC50 (determined as described in Example 7) is 1 rπicromolar or less.
EXAMPLE 2. SYNTHESIS OF 4-METHOXY-N,2,6-TRΪMETHYL-N-{2-[2-(2-METHYL-2-OXIDO-2,8- DlΛZABPlRϋ[4.5]DEC-8-YL)-2-OXOETHOXY]ETHYL}BENZENESULFONAMϊDE (COMPOUND 2)

The title compound is prepared as a yellow oil using the procedure illustrated above. 1H-NMR (400 MHz, CDCl3) & 6.63 (s, 2H), 4.11 (s, 2H), 3.82 (s, 3H), 3.24-3.72 (m, 15H), 2.78 (s, 3H), 2.60
(s, 6H), 1.58-2.58 (m, 6H). LC-MS m/z (M + W): 484.2; retention time: 1.1 min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
EXAMPLE 3. SYNTHESIS OF ADDITIONAL REPRESENTATIVE N-OXIDE ARYL SULFONES AND SULFOXIDES
A. (5R)-4-[(4-METHOXY-2,6-DIMETHYLPHENYL)SULFONYL]-2,2-DIMETHYL-5-({2-[4-( L -METHYL- I - OXIDOPIPERIDIN-4-YL)PIPERAZIN- 1 -YL] -2-OXOETHOX Y}METHYL)MORPHOLINE (COMPOUND 3) rSO2CI

Step I . (3S)-Methyl 4-benzyl-6-(iodomethyl)-6-methy!morphofine-3-carboxylate


The mixture of diastereomers obtained in step 1 is combined with tributyltin hydride (73 mL, 272 mmol) in 400 mL refhixing toluene. AIBN (1 g, 6 mmol) dissolved in 40 mL toluene is added dropwise. After 2 h reflux, the reaction mixture is cooled to it. Saturated aqueous KF solution (800 mL) is added and the resulting heterogeneous mixture is filtered through Celite and the soϋd washed with EtOAc. The combined solutions are transferred to a separatory funnel and washed twice with 200 mL saturated KF, and then once with 20 mL brine. The organic phase is dried over MgSO4, filtered and concentrated in vacuo. The product is purified by flash chromatography eluting with 95/5 hexanes/EtOAc followed by EtOAc to afford (S)-methyi 4-benzyl-6,6-dimethylmorphoIine-3- carboxylate as a clear oil. This oil is dissolved in a mixture of 200 mL MeOH and 20 mL acetic acid. Palladium hydroxide (5 g, 20 wt. % on carbon) is added and the reaction mixture hydrogenated on a Paar apparatus at 50 psi for 10 h. An additional 5 g of palladium hydroxide is added and the hydrogenation is continued at 50 psi for another 25 h. The reaction mixture is filtered through Celite and concentrated in vacuo to afford the title compound. 1H NMR (400 MHz, d-6-DMSO) δ: 3.71-3.74 (m, IH), 3.61-3.65 (m, IH), 3.61 (s, 3H), 3.36 (m, IH), 2.65 (d, IH), 2.46 (d, IH), 1.1 1 (s, 3H)5 1.09 (s, 3H).
Step 3. (S)-Methyl 4-(4-methoxy-2,6-dimethylphenylsulfonyl)-6,6-dimethylmorpholine-3- carboxylate

(S)-Methyl 6,6-dimethylmorpholine-3-carboxyIate (2.3 g, 13 mmol) is dissolved in 50 mL pyridine. 2,6-Dimethyl-4-methoxyphenylsulfonyl chloride (3.1 g, 13 mmol) and DMAP (15 mg) are added. The reaction mixture is stirred at 50 °C for 5 h. The reaction mixture is cooled to it, diluted with 100 mL EtOAc, and transferred to a separatory funnel. The reaction mixture is washed three times with 50 mL water and once with 50 mL brine. The organic phase is dried over MgSO4, fiftered, then concentrated in vacuo. The product is purified by flash chromatography eluting with 8/2 hexanes/EtOAc followed by 1/1 hexanes/EtOAc to afford the title compound.
Step 4. (R)-2-((4-(4-Methoxy-2,6-dimethylphenylsulfonyI)-6,6-dimethyImorpholin-3- yl)methoxy)acetic acid

(S)-Methyl 4-(4-methoxy-2,6-dimethylphenyIsuIfonyl)-6,6-dimethyimorphoIine-3- carboxylate is dissolved in 8 niL of dry THF at 0 0C. DIBAL (8 niL of a IM solution in THF) is added and the reaction mixture stirred for 30 min at 0 0C. An additional 1 mL of DIBAL solution is added and stirring is continued for 30 min. The reaction mixture is quenched by addition of solid Na3SO4 1 IOH2O. The reaction mixture is filtered through Celite and concentrated in vacuo to afford (R)-(4-(4-methoxy-2,6-dimethyiphenylsulfonyi)-6,6-dimethylmorphoIin-3-yl)methanoI. This compound, without further purification, is dissolved in 10 mL of DMA at it. Potassium iodide (20 mg) and sodium hydride (140 mg of a 60% oil dispersion) are added, followed by bromo /m-butyl acetate (0.48 mL, 3.2 mmol). The reaction mixture is stirred for 2 days, and then diluted with 50 mL EtOAc, transferred to a separator/ funnel, and washed once with 20 mL water and once with 20 mL brine. The organic phase is dried over MgSO.*, filtered and concentrated in vacuo. The product is purified by flash chromatography eluting with 9/1 hexanes/EtOAc to afford (R)-teft-butyl 2-((4-(4- methoxy-2,6-dimethylphenylsulfonyl)-6.6-dimethyImoφhoIin-3-yl)methoxy)acetate. This compound is dissolved in 3 mL of 4 M HC! in dioxane, and stirred at 50 0C for 3 h , The reaction mixture is cooled to rt and concentrated in vacuo. The product is purified by flash chromatography, eluting with 1/1 hexanes/EtOAc to afford the title compound.
Step 5. (R)-2-((4-(4-Methoxy-2,6-dimethylphenylsulfonyl)-636-dimethylmoφholin-3-yl)methoxy)-I- (4-(l-methylpiperidin-4-yl)piperazin-l-yI)ethanone

3-y!)methoxy)acetic acid (1 mL of a 0.2 M solution in 95/5 EtOAc/TEA) is added isobutylchloroformate (1 mL of a 0.2 M solution in toluene). The reaction mixture is stirred at rt for 5 min. ] -(3 -Methylpiperidin-4-yl)ρiperazine (1.2mL of a 0.2M solution in toluene) is added and the reaction mixture is stirred for 90 min at rt. The reaction mixture is diluted with 5 mL of EtOAc and washed with 1 mL of IN NaOH. The organic phase is applied to a 10 g silica SPE cartridge and eluted with ΪO mL of EtOAc to remove non-polar impurities, followed by 10 mL of 10/1/1 EtOAc/MeOH/TEA to elute the title compound. The solvent is removed in vacuo to afford the title
compound. 1H NMR (400 MHz, CDCl3) δ: 6.62 (s, 2H), 4.04 (s, 2H), 3.93 (dd, IH)5 3.84-3.86 (m, IH), 3.81 (s, 3H), 3.76 (d, IH), 3.70 (m, IH), 3.56 (m, 3H), 3.47 (m, IH), 3.39 (m, 2H), 2.99 (dd, 2H), 2.90 (d, 2H), 2.60 (s, 6H), 2.52 (m, 4H), 2.25 (s, 3H), 1.95 (t, 2H), 1.75 (d, 2H), 1.56 (m, 2H), 1.15 (s, 3H), l.l l (s, 3H).
Step 6. (5R)-4-[(4-methoxy-2,6-diraethylphenyi)sulfonyl]-2,2-dimethy1-5-({2-[4-(l-methyJ-l- oxidopiperidin-4-yi)piperazm-l-yl]-2-oxoethoxy}methyl)morpholine

A 0.25 M solution of 3-chloroperbenzoic acid in CH2Cl2 (500 μL, 125 μmol) is added to a 0 0C solution of 4-[(4-methoxy-2,6-dimethylphenyl)suIfonyl]-2,2-dimethyl-5-({2-[4-(I - methy!piperidin-4-yI)piperazin-l-yl]-2-oxoethoxy}methyl)moφholine (58.7 mg, 104 μmol) in CH2Cl2 (500 μL). After 1.5 Ii , the volatiies are removed under reduced pressure. The resulting residue is purified by thin layer chromatography (2% NH4OH in 4 : 1 CH2Cl2 : MeOH) to afford the title compound as a tan solid. 1H-NMR (400 MHz, CDCl3) & 6.62 (s, 2H), 4.06 (s, 2H), 3.69-3.96 (m, 4H), 3.82 (s, 3H), 3.56-3.62 (m, 3H), 3.40-3.45 (m, 2H), 3.29-3.36 (m, 2H), 3.22 (s, 3H), 3.12-3.22 (m, 2H)5 2.93-3.02 (m, 2H), 2.60 s, 6H), 2.38-2.60 (m, 7H), 1.64-1.71 (m, 2H), 1.15 (s, 3H), 1.10 (s, 3H). LC-MS m/∑ (M + H+): 582.3. Retention time: I.I min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
B. { l-[({(3R)-4-[(4-METHOXY-2,6-DIMETHYLPHENYL)SULFONYL]-6,6-DIMETHYLMORPHOLIN-3- YL}METHOXY)ACETYL]PiPERIDIN-4-YL}DIMETHYLAMlNE OXIDE (COMPOUND 4)

The title compound as a pale yellow film is prepared as indicated above. 1H-NMR (400 MHz, CDCl3) & 6.62 (s, 2H), 4.70-4.82 (m, IH), 3.95-4.33 (m. 3H), 3.53-3.94 (m, 4H), 3.80 (s, 3H), 3.27- 3.45 (m, IH), 3.16 (s, 3H), 3.13 (s, 1.5H), 3.1 1 (s, 1.5H), 2.86-3.08 (m, 3H), 2.24-2.67 (m, 4H), 2.58
(s, 6H), 1.52-1.82 (m, 2H), 1.12 (s, 3H), 1.05 (s, 1.5H), 0.97 (s, 1.5H). LC-MS m/z (M + H+): 527.3. Retention time: 1.1 min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
C. (2-{ L -[( {(3R)-4-[(4-METHOXY-2!6-DIMETHYLPHENYL)SULFONYL]-6,6-DIMETHYLMORPHOLIN-3- YL}METHOXY)ACETYL]PIPERIDIN-4-YL}ETHYL)DIMETHYLAMINE OXIDE (COMPOUND 5)

The title compound as a pale yellow film is prepared as indicated above. 1H-NMR (400 MHz, CDCl3) & 6.62 (s, 2H), 4.52 (dd, IH), 3.52-4.19 (m, 8H), 3.63 (ss 3H), 3.23-3.32 (m, 2H), 3.18 (s, 1.5H), 3.16 (s, 3H), 3.14 (s, 1.5H), 2.81-3.05 (m, 2H)5 2.59 (s, 6H), 2.48-2.61 (m, 2H), 1.52-2.01 (m, 5H), 1.10-1.35 (m, 2H), 1.14 (s, 3H), LlO (s, 1.5H), 1.04 (s, 1.5H). LC-MS m/z (M + H+): 556.2. Retention time: 1.1 min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
D. N-[2-(2-{9-[(DlMETHYLNITRORYL)METHYL]-3-AZASPIRO[5.5]UNDEC-3-YL}-2- OXOETHOXY)ETHYL]-4-METHOXY-N,2,6-TRIMETHYLBENZENESULFONAMIDE (COMPOUND 6)

The title compound as a pale yellow film is prepared as indicated above. 1H-NMR (400 MHz, CDCl3) δ: 6.61 (s, 2H), 4.09 (d, 2H), 3.80 (s, 3H), 3.66 (t, 2H), 3.46-3.55 (m, 2H), 3.28-3.40 (m, 4H), 3.20 (s, 6H), 3.14 (t 2H), 2.77 (s, 3H), 2.59 (s, 6H), 2.00-2.14 (m, I H), 1.79-1.88 (m, 2H), 1.64-3.72 (m, 2H), 1.44-1.51 (m, 2H), 1.20-1.36 (m, 6H). LC-MS m/z (M + H'): 540.3; retention time: 1.1 min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
E. 4-METHOXY_N,2,6-TRIMETHYL-N-(2-{2-[4-(l -METHYL- l-OXIDOPIPERIDIN-4- YL)PIPERAZIN-1 -
YL] -2-OXOETHOXY} ETHYL)BENZENESULFONAMIDE (COMPOUND 7)

The title compound as a colorless solid is prepared as indicated above. 1H-NMR (400 MHz, CDCl3) δ: 6.61 (s, 2H), 4.09 (d, 2H), 3.79 (s. 3H), 3.66 (t, 2H), 3.58 (brs, 2H), 3.39-3.43 (m, 6H),
3.16-3.22 (m, 4H), 2.76 (s, 3H), 2.74 (m, 3H), 2.57 (s, 6H), 2.42 (brs, 2H), 1.68 (brs, 2H). LC-MS m/z
(M + H+): 513.3; retention time. 1.1 miπ. The IC50 (determined as described in Example 7) is I micromolar or less.
F. 2,6-DLCHLORO-N-METHYL-N-(2- {2-[4-( L -METLIYL- L -OXIDOPIPERIDIN-4-YL)PIPERAZIN-L -YL]-2- OXOETHOXY} ETHYL)BENZENESULFONAMIDE (COMPOUND 8)

The title compound as a colorless solid is prepared as indicated above. 1H-NMR (400 MHz,
CDCl3) S: 7.44 (d, 2H), 7.30 (t, IH), 4.15 (s, 2H), 3.72-3.75 (m, 2H), 3.60 (brs, 2H), 3.53-3.55 (m, 2H), 3.46 (brs, 3H), 3.36 (d, 2H), 3.15-3.21 (m, 4H), 3.01 (s, 2H), 2.58-2.61 (m, 4H), 2.46-2.50 (m,
4H)5 1.69 (d, 2H). LC-MS m/z (M + H~): 523.2; retention time: 1.0 min. The IC50 (determined as described in Example 7) is 1 micromolar or less.
EXAMPLE 4. ADDITIONAL REPRESENTATIVE N-OXIDE ARYL SULFONES AND SULFOXIDES
Using routine modifications, the starting materials may be varied and additional steps employed to produce other compounds provided herein. Compounds listed in Table I are prepared using such methods.
Table I
Representative N-Oxide Aryl Sulfones and Sulfoxides


EXAMPLE 5. PREPARATION OF BrTRANSFECTED CELLS
This Example illustrates the preparation of Bj -transfected cells for use in Bi binding and modulation assays (Examples 6 and 7).
Cynomolgus macaque lung total RNA is isolated as described by Chomzynski et al. (1987) Anal. Biochem. 762: 156-159. A cDNA encoding Bj is cloned from the total RNA by reverse transcriptase-polymerase chain reaction (RT-PCR) with the following oligonucleotides:
Primer 1: GGCGCTAGCCACCATGGCATCCTGGCCCCCTC (SEQIDNO:!)
Primer 2: AGCCGTCCCAGATCTGAAC (SEQIDNO:2)
Primer 3: GATCTGGGACGGCTTGGATG (SEQIDNO:3) Primer 4: CGGAGCTCTTAATTCCGCCAGAAAAGTTGGA (SEQΪDNO:4)
Primer pairs 1 & 2 and 3 & 4 are used to generate overlapping cDNA fragments corresponding to the entire protein coding sequence of cynomolgus macaque Bj cDNA are isolated and linked to form the full-length coding sequence (GenBank Accession Number AY788905), The construct is cloned into pcDNA 3.1 (Invitrogen, Carlsbad, CA) and transfected into Chinese hamster ovary (CHO) ceils using Lipofectamine (Invitrogen), resulting in cynomolgus macaque Brexpressing CHO cells. Alternatively, the construct is cloned into pBAKPAK9 (Clontech, Mountain View, CA) and transfected into Sf9 cells to generate clonal baculovirus stocks. Clonal cell lines stably expressing the cynomolgus macaque Bi receptor are selected in G418. A single clonal line that exhibits high levels of receptor expression is chosen for use in binding and calcium mobilization assays (Examples 6 and
7). Clonal baculovirus stocks are used to infect Sf9 cells such that the infected cells express high levels of recombinant Bi receptors. These cells are used in radioligand binding assays (Example 6).
EXAMPLE 6. Bj RECEPTOR BINDING ASSAYS This Example illustrates a representative Bi receptor binding assay that may be used to determine the binding affinity of compounds for B,.
A. [3H]-DESARG10KALLIDIN BINDING TO INTACT IMR-90 CELLS OR CHO CELLS STABLY EXPRESSING RAT Bi
IMR-90 cells, which endogenously express human Bi, are seeded into 24 well plates at 65,000 cells per well, cultured overnight, and then treated for 3 h with 0.2 ng/mL interleukin-1 beta to induce B. expression (Menke, et ai, (1994) J. Biol Chem. 269:21583-86). CHO cells stably expressing rat B5 are seeded into 24 well plates at 200,000 cells per well and cultured overnight. The cells are then washed 3 times with phosphate buffered saline (PBS). One hundred fifty microliters of binding buffer (50 mM Tris 7.4, 0.14 mg/mL, bacitracin, and 1 mg/mL BSA) is added to each well. Various concentrations of test compound are added to each well from DMSO solutions such the final DMSO concentration is 1% by volume; some wells receive DMSO only, and some wells receive DMSO plus 10 μM desArg10KalIidin to define non-specific binding. All wells then receive 0.3 nM (final concentration) [3H]-desArg10Kallidin. The plates are allowed to sit for 2 h at room temperature. Cells are then washed three times, and lysed with 400 μl Ultima Gold scintillation fluid (PerkinElmer; Boston, MA; 20 min incubation). The fluid is then transferred to counting vials counted in a Packard liquid scintillation counter (PerkinElmer). The number of counts present in the scintillation fluid is plotted as a function of antagonist compound concentration and fitted to a logistical equation using SigmaPlot (Systat Software, Point Richmond, CA) to determine each compound's IC50 and K1 (e.g., as described by Szallasi. et al. (1993) J. Pharmacol Exp. Ther. 25(5:678-83).
B. [3 H]-DESARD!0KALLIDIN BINDING TO MEMBRANE HOMOGENATES OF SF9 CELLS EXPRESSING
CYNOMOLGUS MACAQUE B ,
Sf9 cells infected with a baculovirus carrying the coding sequence for cynomolgus macaque Bi are harvested by centrifugation and frozen at -SO 0C. Pellets are subsequently resuspended on ice in Tris buffered saline (TBS; 50 mM Tris (pH 7.4), 120 mM NaCl), and cells are homogenized using a polytron for 30 seconds. The crude membrane fraction is collected by centrifugation at 20,000 rpm. Membranes are washed two times with TBS and collected by centrifugation each time. Protein content of the membranes is determined after the last wash and the concentration is adjusted to 0.7 μg/uL with binding buffer (50 mM Tris 7.4, 0.14 mg/mL bacitracin, and 1.0 mg/mL BSA). To perform the binding assay, 150 microliters of membrane fraction is added to each well of a 96-well plate along with 50 μl [3H]-desArg!0Kallidin (0.3 nM final) and test compound in DMSO (final
DMSO concentration = 1%). Some wells receive DMSO only, and some wells receive DMSO plus 10 μM desArg10Kallidin to define non-specific binding. The 96 well plates are allowed to sit for 2 h at room temperature. Membrane proteins are then harvested by filtration onto GF/C filtermats (PerkinElmer) pre-soaked for 1 hr in 0.5 % polyethylenimine. After filtration, filters are dried and then counted in a Beta plate counter. The number of counts present in the scintillation fluid is plotted as a function of antagonist compound concentration and Fitted to a logistical equation using SigmaPlot (Systat Software, Point Richmond. CA) to determine each compound's IC50 and K1 (e.g., as described by Szallasi, et al. (1993) J. Pharmacol. Exp. Thβr. 266:678-83).
EXAMPLE 7. CALCIUM MOBILIZATION ASSAY This Example illustrates representative calcium mobilization assays for use in evaluating test compounds for agonist and antagonist activity.
Cynomolgus macaque Brexpressing CHO cells (Example 5) are plated in a 96 well plate. The cells are cultured for 1 day, after which culture media is emptied from the plate and replaced with 50 μl of KRH (Krebs-Ringer HEPES buffer: 25 mM HEPES, 5 mM KCl3 0.96 mM NaH2PO4, 1 mM MgSO4, 2 mM CaCl2, 5 mM glucose, 1 rnM probenecid, pH 7.4) supplemented with the calcium- sensitive fluorescent dye Fluo4-AM (5 μg/ml; Teflabs, Austin, TX). The cells are then incubated at 37 0C in an environment containing 5% CO2. After the 1 hour incubation, the dye solution is removed from the plate, the plate is washed once with KRH, and 100 μL KRH is added.
DETERMINATION OF B, AGONIST EC50 100 μL KRH + 2% DMSO is added to each well of cells, such that the final volume in each well is 200 microliters and the final DMSO concentration is 1%. Various concentrations of the Bi agonist desArg!°Kallidin are added. Addition of desArg 10KaI lid in elicits a fluorescent response as consequence of increased intracellular calcium. This response is measured with a FLIPR instrument (Molecular Devices, Sunnyvale, CA) and determined to be desArg10Kallidin concentration dependent. A plot of maximum fluorescent response as a function of desArg10Kalligin is generated and an EC50 (concentration required to elicit a 50% of maxima! response) for the response is determined using the equation: y=a*(l/(l+(b/x)c))
In this equation, y is the maximum fluorescence signal, x is the concentration of the B, agonist, a is the Emax, b corresponds to the EC50 value and c is the Hill coefficient.
By replacing the desArg10Kallidin with a test compound, this assay is also used to assess Bi agonist activity of the test compound.
DETERMINATION OF ANTAGONIST ACTIVITY
Various concentrations of test compounds are added to the cell plate prepared as described above in 100 μL KRH + 2% DMSO, such that the final volume in each well is 200 microliters and the final DMSO concentration is 1%. The EC50 concentration of desArg10Kaliidin is then added to each well of plates containing test compound to determine the extent to which each test compound inhibits an agonist-induced Bi response. The maximum fluorescent response is plotted as a function of test compound concentration in order to determine the IC50 (concentration required to inhibit 50% of the effect of agonist) for each compound at B] . Antagonists of B; decrease this response by at least about 20%, preferably by at least about 50%, and most preferably by at least 80%, as compared to matched control (i.e., cells treated with desArg!°Kallidin at the EC50 concentration in the absence of test compound), at a concentration of I O micromoiar or less, preferably 1 micromolar or less.
Alternatively, the data is analyzed as follows. First, the average maximum relative fluorescent unit (RFU) response from negative control wells (no agonist) is subtracted from the maximum response detected for each of the other experimental wells. Second, average maximum RFU response is calculated for the positive control wells (agonist wells). Then, percent inhibition for each compound tested is calculated using the equation:
P nercen *t τ Inh i i -Ubi+ti-on = i 1n0n0 - i 1n0n0 *
 f "I
L Peak signal in Agonist Wells J
The % inhibition data is plotted as a function of test compound concentration and test compound IC5O is determined using a linear regression in which x is ln(concentration of test compound) and y is ln(ρercent inhibition/( 100 - percent inhibition). Data with a percent inhibition that is greater than 90% or less than 15% are rejected and are not used in the regression. The IC50 is
(-intercepϊ7slope) e
EXAMPLE 8. MDCK CYTOTOXICITY ASSAY
This Example illustrates the evaluation of compound toxicity using a Madin Darby canine kidney (MDCK) cell cytotoxicity assay. 1 μL of test compound is added to each well of a clear bottom 96-wel! plate (Packard,
Meriden, CT) io give final concentration of compound in the assay of 10 μM, 100 μM or 200 μM.
Solvent without test compound is added to control wells.
MDCK cells, ATCC no. CCL-34 (American Type Culture Collection, Manassas, VA), are maintained in sterile conditions following the instructions in the ATCC production information sheet. Confluent MDCK cells are trypsinized, harvested, and diluted to a concentration of 0.1 x 10⁶ cells/mL with warm (37°C) medium (VITACELL Minimum Essential Medium Eagle, ATCC catalog # 30-
2003). 100 μL of diluted cells is added to each well, except for five standard curve control wells that contain 100 μL of warm medium without cells. The plate is then incubated at 370C under 95% O2,
5% CO2 for 2 hours with constant shaking. After incubation, 50 μL of mammalian cell lysis solution (from the Packard (Meriden, CT) ATP-LITE-M Luminescent ATP detection kit) is added per well, the wells are covered with PACKARD TOPSEAL stickers, and plates are shaken at approximately 700 rpm on a suitable shaker for 2 min. Compounds causing toxicity will decrease ATP production, relative to untreated cells. The
ATP-LITE-M Luminescent ATP detection kit is generally used according to the manufacturer's instructions to measure ATP production in treated and untreated MDCK cells. PACKARD ATP LITE-M reagents are allowed to equilibrate to room temperature. Once equilibrated, the lyophilized substrate solution is reconstituted in 5.5 mL of substrate buffer solution (from kit). Lyophilized ATP standard solution is reconstituted in deionized water to give a 10 mM stock. For the five control wells, 10 μL of serially diluted PACKARD standard is added to each of the standard curve control wells to yield a final concentration in each subsequent well of 200 nM, 100 nM, 50 nM, 25 nM, and 12.5 nM. PACKARD substrate solution (50 μL) is added to all wells, which are then covered, and the plates are shaken at approximately 700 rpm on a suitable shaker for 2 min. A white PACKARD sticker is attached to the bottom of each plate and samples are dark adapted by wrapping piates in foil and placing in the dark for 10 min. Luminescence is then measured at 22°C using a luminescence counter (e.g., PACKARD TOPCOUNT Microplate Scintillation and Luminescence Counter or TECAN SPECTRAFLUOR PLUS), and ATP levels calculated from the standard curve. ATP levels in cells treated with test compound(s) are compared to the levels determined for untreated cells. Cells treated with 10 μM of a preferred test compound exhibit ATP levels that are at least 80%, preferably at least 90%, of the untreated cells. When a 100 μM concentration of the test compound is used, cells treated with preferred test compounds exhibit ATP levels mat are at least 50%, preferably at least 80%, of the ATP levels detected in untreated cells.
Claims
What is claimed is:
1. A compound of the formula:
Ar O=S=O O Q-
RY RB RD or a pharmaceutically acceptable salt or hydrate thereof, wherein: A is N, C or CH;
== represents a single or double bond; Ar is phenyl or a 5- or 6-membered heteroaryl, each of which is substituted with from 0 to 4 substituents independently chosen from Rj; Rx and Rγ are:
(i) independently chosen from:
(a) hydrogen; and
(b) C|-C6alkyi, C^-C^alkenyl, C2-C6alkynyl, C|-C6haloalkyl, and (CrCscycloalky!)CrC4alkyL each of which is substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, amino and C , -C4 alky 1; or
(ii) taken together to form a 4- to 10-membered carbocycle or heterocycle that is substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, halogen, amino and Cr Qalkyl;
Y is a group of the formula (CH2VQ-(CH2)P, which is substituted with from 0 to 4 substituents independently chosen from (i) amino, hydroxy, cyano, C,-C6aikyl, C2-C6aikenyl, C2-C6alkynyl and CrC6haloalkyl; (ii) substituents of the same carbon atom or adjacent carbon atoms that are taken together to form Cs-Qcycloalkyl, and (iii) substituents that are taken together with Rx or Rγ to form a 4- to 7-membered carbocycle or heterocycle. wherein:
Q is absent, CH3, CH=CH, O, S or NRi o, wherein R]0 is hydrogen or C]-C6alkyl; and r and p are independently chosen integers ranging from 0 to 6;
W is C]-C6alkylene that is substituted with from 0 to 4 substituents independently chosen from R20; Z is absent, divalent phenyl, NR0 or CRERF wherein:
RG is hydrogen, Cj -Chalky 1 or taken together with RB or RD to form a 4- to 7-membered heterocycloalkyl that includes W and is substituted with from 0 to 4 substituents independently chosen from R2o;
RE is hydrogen, Ci-Cβalkyl or taken together with RB to foπn a 4- to 7-membered heterocycloalkyl that includes W and is substituted with from 0 to 4 substituents independently chosen from R20; and
RF is hydrogen, Cj-Cealkyl or taken together with RD to foπn a 4- to 7-membered heterocycloalkyl that includes V and is substituted with from 0 to 4 substituents independently chosen from R20; V is absent or CrC6alkylene that is substituted with from 0 to 4 substituents independently chosen from (i) R2O and (ii) groups that are taken together with RD to form a 4- to 7-membered heterocycle that is substituted with from 0 to 4 substituents independently chosen from R20; Each Ri is independently chosen from:
(i) halogen, hydroxy, cyano, amino, nitro, aminocarbonyl, aminosulfonyl and -COOH;
(ii) CrQalky], C2-C6alkenyl, C3-C6alkynyl, (C3-C8cycloa!ky])C0-C4alkyl, C1-C6alkoxy, Cr Qalkylthio, Ci-Cόalkylsulfinyl, Cj-Cealkoxycarbonyl, Ci-C6alkylsulfonyIC0-C4a!kyl, mono- or di-(Ci-C6alkyl)aminoCo-C4alkyl, mono- or di-(Ci-C6alkyI)aminosulfonylCo-C4alkyl, mono- or di-(C]-C6alkyl)aminocarbonylCo-C4alkyl, and (4- to 8-membered heterocycloalky I)C0- C4alkyl; each of which is substituted with from 0 to 6 substituents independently chosen from halogen, hydroxy, cyano and amino; and
(iii) groups that are taken together to form a fused carbocyclic ring that is substituted with from 0 to 4 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, Q- Cealkyl, Cj-Cβhaloalkyl, Ci-Cealkoxy, and mono- or di-(Ci-Cόalkyl)aminoCo-C4alkyi; RB is:
(i) hydrogen, Q-Qalkyl, C2-Qalkenyl, C2-Qa!kynyl, CpQhaloalkyl, (C3-C8cycIoaIkyl)Cr C4alkyl, C2-Cgalkyl ether, mono- or di-(CrC6alkyl)aminoQ-C4alkyl, or (4- to 7-membered heterocycloalkyl)Co-C4alkyl, each of which is substituted with from 0 to 6 substituents independently chosen from:
(a) amino, halogen, hydroxy, cyano and oxo; and
(b) Ci-Qalkyl, (C3-C3cycloalkyI)C0-C4alkyl, Q-Qalkoxy, phenylC0-C4alkyl and (5- or 6- membered heterocycIe)Co-C4alkyl, each of which is substituted with from 0 to 4 substituents independently chosen from amino, cyano, halogen, hydroxy, C 1 -Chalky 1, (C3- Cscycloalkyl)Co-C4alkyl, Cj-Cehaloalkyl, CrCsaikoxy, Ci-C6alkoxycarbonyi, mono- or di-(C]-C6alkyl)amino, pheny!Co-C4aIkyl and phenylC0-C4alkoxy; or
(ii) taken together with RE or R0 to form an optionally substituted heterocycloalkyl; Rc is absent, Ci -Chalky 1 or taken together with RD to from a 4- to 7-membered heterocycle that is substituted with from 0 to 4 substituents independently chosen from R20; RD is Ci-Cβaikyl or taken together with Rc, RF, RG or a substituent of V to form an optionally substituted 4- to 7-membered heterocycle; and Each R20 is independently selected from:
(i) hydroxy, oxo, cyano and amino; and
(ii) Ci-C6alkyl, Q-C6a!koxy, mono- or di-(C1-C6alkyl)ammoC0-C4alky]J (C3 -Q ocarbocyc Ie)C0- C4alkyi and (4- to 10-membered heterocycie)C0-C4alkyl, each of which is substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, Q- Qalkyl, Q-Q,alkoxy, (C3-C!0carbocyc]e)Co-C4alkyl and (4- to 10-membered heterocyc Ie)Co- Qaϊkyl.
2. A compound or salt or hydrate thereof according to claim 1, wherein Ar is phenyl, naphthyl, pyridyl or pyrimidinyl, each of which is substituted with from 0 to 4 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, Q-C6alkyl, Ci-C6haloalkyl, Q- Cgalkoxy, and mono- or

3. A compound or salt or hydrate thereof according to claim 1, wherein Ar is phenyl that is substituted with from 0 to 4 substituents independently chosen from R\.
4. A compound or salt or hydrate thereof according to claim 2, wherein Ar is phenyl that is substituted with 1, 2 or 3 substituents independently chosen from halogen, hydroxy, cyano, amino, nitro, CrC6alkyl, Q-C6haloalkyl, Q-C6alkoxy, Q-C6haloalkoxy a"d mono- or di-(Q- C6alkyl)am in oC0-C4aI ky 1.
5. A compound or salt or hydrate thereof according to claim 1, wherein each substituent represented by Rj is independently chosen from halogen, hydroxy, Q-Q,alkyl, Q-Qhafoaikyl, Q- Qalkoxy and Q-C6haloalkoxy.
6. A compound or salt or hydrate thereof according to claim 1, wherein Ar is

7. A compound or salt or hydrate thereof according to claim 6, wherein R5 is methyl or methoxy and R4 and Rg are each methyl.
8. A compound or salt or hydrate thereof according to any one of claims 1-7, wherein the sum of r and p ranges from 1 to 6.
9. A compound or salt or hydrate thereof according to claim 8, wherein p is not zero.
10. A compound or salt or hydrate thereof according to any one of claims 3 -9, wherein Y is a group of the formula (CH2)rG-(CH2)p that is substituted with from 0 to 4 substituents independently chosen from amino, hydroxy, cyano, Ci-C6alkyl, Q-Cβalkenyl, C2-C6alkynyl, Ci-C6haloalkyl and substituents of the same carbon atom or adjacent carbon atoms that are taken together to form C3- C6cycloaikyl.
11. A compound or salt or hydrate thereof according to claim 10, wherein Y is -G-CH2-, -CH2-O-CH2-, -0-CH2-CH2-, -CH2-O-CH2-CH2- Or -CH2-CH2-O-CH2-.
12. A compound or salt or hydrate thereof according to claim 1 1 , wherein Y is -0-CH2-.
13. A compound or salt or hydrate thereof according to any one of claims 1-9, wherein Y is- N(R1O)-CH2-, -CH2-CH2- -CH=CH- or -CH2-CH2-CH2-.
14. A compound or salt or hydrate thereof according to any one of claims 1-13, wherein Rx and Ry are independently chosen from hydrogen and Ci-C6alkyl.
15. A compound or salt or hydrate thereof according to anyone of claims 1-14, wherein the compound has the formula:
Ar O=S=O O 0-
RY RB RD wherein Rx and Ry are:
(i) independently chosen from:
(a) hydrogen; and
(b) CrCήalkyl, C2-C6alkenyl, C2-C6alkynyl, CrC6haloalkyl, and (C3-Cscycloalkyl)C0-C4alkyl, each of which is substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, amino and C]-C4alkyl; or
(ii) taken together to form a 4- to 1 0-membered heterocycle that is substituted with from 0 to 4 substituents independently chosen from oxo, hydroxy, halogen, amino and CrC8alkyi.
16. A compound or salt or hydrate thereof according to claim 15, wherein Rx and Rγ are taken together to form an optionally substituted 4- to 8-membered heterocycloalkyl.
17. A compound or salt or hydrate thereof according to claim 16, wherein the compound satisfies the formula:

X is CH2, NR3, O, S, SO or SO2; m and q are independently O, 1 or 2;
R2 represents from O to 4 substituents independently chosen from oxo, hydroxy and Cs-C6alkyl; and
R3 is hydrogen, Ci-Qalkyl or C|-Qalkanoyl.
IS. A compound or salt or hydrate thereof according to claim 17, wherein the compound satisfies the formula:
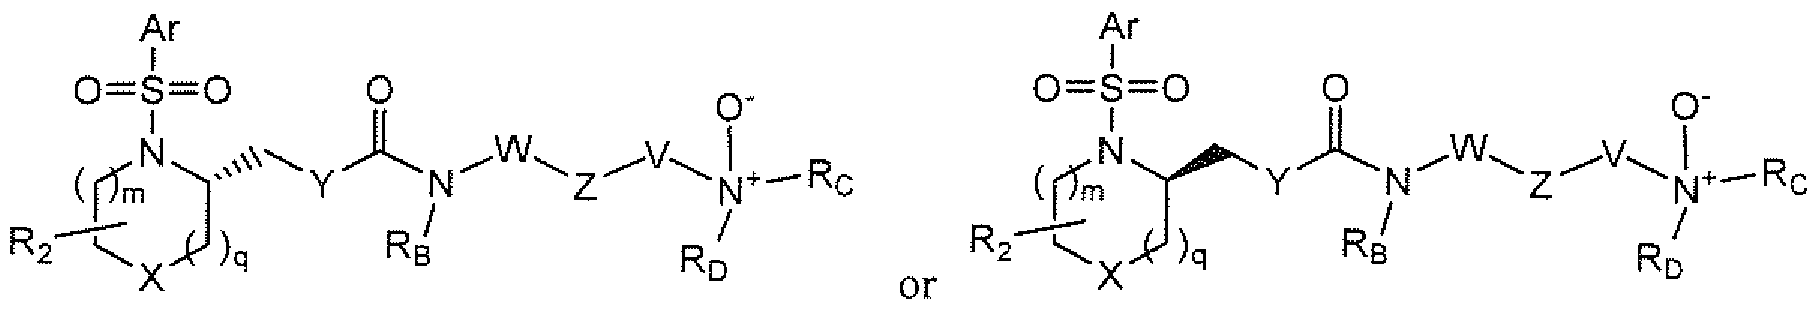
19. A compound or salt or hydrate thereof according to claim 17 or claim 18, wherein X is CH2.
20. A compound or salt or hydrate thereof according to claim 17 or claim 18, wherein X is O.
21. A compound or salt or hydrate thereof according to claim 17 or claim 18, wherein:

22. A compound or salt or hydrate thereof according to any one of claims 17-21, wherein R2 represents from 0 to 4 substituents independently chosen from CrC2alkyl.
23. A compound or salt or hydrate thereof according to claim 22, wherein R2 represents gem-dimethyl.
24. A compound or salt or hydrate thereof according to any one of claims 1-23, wherein Z is NRG or CRERF.
25. A compound or salt or hydrate thereof according to claim 24, wherein RE is taken together with RB to form a 4- to 7-membered heterocycloalkyl that is substituted with from 0 to 4 substituents independently chosen from: (i) hydroxy, oxo, cyano and amino; and
(ii) Ci-Cealkyl, Ci-C6alkoxy, mono- or di-(Ci-C6alkyI)aminoCo-C4alkyl, (C3-Ci0carbocycle)Co- C4alkyl and (4- to 10-membered heteiOcycle)C0-C4aIkyl; each of which is substituted with from 0 to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, C1- C6aϊkyl, Ci-C6aikoxy, (C3-Ciocarbocycle)Co-C4alkyl and (4- to 10-membered heterocycle)C0- C4alkyl.
26. A compound or salt or hydrate thereof according to claim 25, wherein the compound satisfies the formula:

RD is C i -Chalky 1; and
V is absent or C[-C4alkylene.
27. A compound or salt or hydrate thereof according to claim 26, wherein the compound satisfies the formula:

X is NR3, O, S, SO or SO2; and
R2 represents from 0 to 4 substituents independently chosen from oxo, hydroxy and Ci-Qalkyl.
28. A compound or salt or hydrate thereof according to claim 25, wherein the compound satisfies the formula:

RD is Ci-C6alkyl; and
V is CrQalkyiene.
29. A compound or salt or hydrate thereof according to claim 28, wherein the compound satisfies the formula:

X is NR3, O, S, SO or SO2; and
R2 represents from O to 4 substituents independently chosen from oxo, hydroxy and Ci-Cgalkyl.
30. A compound or salt or hydrate thereof according to claim 26 or claim 28, wherein Rx and Ry are independently hydrogen or Ci-Cgalkyl.
31. A compound or salt or hydrate thereof according to claim 25, wherein the compound satisfies the formula:

32. A compound or salt or hydrate thereof according to claim 31, wherein the compound satisfies the formula:

X is NR3, O, S, SO or SO2; and
R2 represents from O to 4 substituents independentiy chosen from oxo, hydroxy and Ci -Cg alky L
33. A compound or salt or hydrate thereof according to claim 31, wherein Rx and Ry are independently hydrogen or Ci -Chalky I.
34. A compound or salt or hydrate thereof according to claim 24, wherein Z is CRERF and Rp is taken together with RD to form a 4- to 7-membered heterocycloalkyl that is substituted with from O to 4 substituents independentiy chosen from:
(i) hydroxy, oxo, cyaπo and amino; and
(ii) CrC6alkyl, Ci-C6alkoxy, mono- or di-(Ci-Cfialkyi)aminoCo-C4alkyl, (C3-C10carbocyc!e)Co- C4alkyl and (4- to 10-membered heterocycle)Co-C4alkyl; each of which is substituted with from O to 2 substituents independently chosen from hydroxy, halogen, oxo, cyano, Cr C6alkyl, Ci-C6aikoxy, (C3-Ciocarbocycle)C0-C4alkyl and (4- to 10-membered heterocycle)C0- C4alkyl.
35. A compound or salt or hydrate thereof according to claim 34, wherein the compound satisfies the formula:

36. A compound or salt or hydrate thereof according to claim 35, wherein Rx and Rγ are independently hydrogen or C]-Cfialkyl.
37. A compound or salt or hydrate thereof according to claim 35, wherein the compound satisfies the formula:

X is NR3, O, S, SO or SO2; and
R2 represents from O to 4 substituents independently chosen from oxo, hydroxy and Ci-Cgalkyl.
38. A compound or salt or hydrate thereof according to claim 24, wherein Z is NR0 and RG is hydrogen or Ci-C6alkyl.
39. A compound or salt or hydrate thereof according to claim 1, wherein the compound is:
4-Methoxy-N72,6-trimethyl-N-{2-[2-(9-methyl-9-oxido-3,9-diazaspiro[5.5]undec-3-yl)-2- oxoethoxt]ethyl}benzenesulfonamide;
4-Methoxy-N,2,6-trimethyl-N-{2-[2-(2-methyI-2-oxido-2,8-diazaspiro[4.5]dec-8-yl)-2- oxoethoxy]ethyl}benzenesulfonamide;
(5R)-4-[(4-Methoxy-2,6-dimethylphenyl)sulfonyl]-2,2-dimethyl-5-({2-[4-(l -methyl- 1-oxidopiperidin-
4~yl)piperazin-l-yl]-2-oxoethoxy}methyl)morphoHne;
{ l-[({(3R)-4-[(4-methoxy-2,6-dimethylphenyi)suifonyl]-6,6-dimethyimoφholin-3- yl}methoxy)acetyl]piperidin-4-yI}dimethylamine oxide;
(2-{ l-[({(3R)-4-[(4-methoxy-2,6-dimethylphenyl)sulfonyI]-6,6-dimethyImorphoIin-3- yl}methoxy)acetyI]piperidin-4-yi}ethyl)dimethylamine oxide;
N-[2~(2-{9-[(Dimethyinitroryl)methyl]-3-azaspiro[5.5]undec-3-yl}-2-oxoethoxy)ethyi]-4-methoxy- N,2,6-trimethylbenzenesulfonamide;
4-Methoxy_N,2,6-trimethyI-N-(2-{2-[4-(] -methyl- 1 -oxidopiperidm-4-y])piperazin- 1 -yFj-2- oxoethoxy} ethyl)benzenesulfonamide; or
2,6-Dichloro-N-methyl-N-(2-{2-[4-(l-methyl-l-oxidopiρeridin-4-yl)piperazin-I-y1]-2- oxoethoxy } ethy l)benzene su I fonamide .
40. A compound or salt or hydrate thereof according to any one of claims I -39wherein the compound exhibits no detectable agonist activity an in vitro assay of Bi agonism.
41. A compound or salt or hydrate thereof according to any one of claims l-40wherein the compound has an IC50 value of I micromolar or less in an in vitro assay of B] antagonism.
42. A pharmaceutical composition, comprising at least one compound or salt or hydrate thereof according to any one of claims 1-41 in combination with a physiologically acceptable carrier or excipient.
43. A pharmaceutical composition according to claim 42 wherein the composition is formulated as an injectible fluid, an aerosol, a cream, a gel, a pill, a capsule, a syrup or a transdermal patch.
44. A method for inhibiting induction of agonist-induced Bi activity in vitro, the method comprising contacting B] receptor with at least one compound or salt or hydrate thereof according to any one of claims 1-41 under conditions and in an amount sufficient to detectably inhibit agonist- induced Bj activity.
45. A method for inhibiting induction of agonist-induced Bi activity in a patient, comprising contacting cells expressing B1 receptor with at least one compound or salt or hydrate thereof according to any one of claims 1-41 , in an amount sufficient to detectably inhibit agonist- induced B] activity in cells expressing a cloned Bj receptor in vitro, and thereby inhibiting agonist- induced Bi activity in the patient.
46. A method according to claim 45, wherein the patient is a human.
47. A method for treating a condition responsive to Bi receptor modulation in a patient, comprising administering to the patient a therapeutically effective amount of at least one compound or salt or hydrate thereof according to any one of claims 1-41. and thereby alleviating the condition in the patient.
48. A method according to claim 47, wherein the condition is inflammation or pain.
49. A method according to claim 47, wherein the condition is cough, asthma, vascular edema or epilepsy.
50. A method for treating pain in a patient, comprising administering to a patient suffering from pain a therapeutically effective amount of at least one compound or salt or hydrate thereof according to any one of claims 1-41, and thereby alleviating pain in the patient.
51. A method according to claim 50, wherein the patient is suffering from inflammatory pain, acute pain, dental pain, back pain, surgical pain, headache, neuropathic pain or pain from osteoarthritis or trauma.
52. A method according to claim 50, wherein the patient is a human.
53. A compound or salt or hydrate thereof according to claim I, wherein the compound is radiolabeled.
54. A method for determining the presence or absence of Bi receptor in a sample, comprising the steps of:
(a) contacting a sample with a compound or salt or hydrate thereof according to any one of claims 1-41, under conditions that permit binding of the compound to Bi receptor; and
(b) detecting a signal indicative of a level of the compound bound to Bi receptor, and therefrom determining the presence or absence of Bi receptor in the sample.
55. A method according to claim 54, wherein the compound is radiolabeled, and wherein the step of detection comprises the steps of:
(i) separating unbound compound from bound compound; and
(ii) detecting the presence or absence of bound radiolabel in the sample.
56. A packaged phaπnaceutical preparation, comprising:
(a) a pharmaceutical composition according to claim 42 in a container; and
(b) instructions for using the composition to treat pain.
57. The use of a compound or salt or hydrate thereof according to any one of claims 1-41 for the manufacture of a medicament for the treatment of a condition responsive to B| receptor modulation.
58. A use according to claim 57, wherein the condition is an inflammatory condition or pain.
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US82326506P | 2006-08-23 | 2006-08-23 | |
| US60/823,265 | 2006-08-23 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2008024692A1 true WO2008024692A1 (en) | 2008-02-28 |
Family
ID=39107134
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/US2007/076266 Ceased WO2008024692A1 (en) | 2006-08-23 | 2007-08-20 | N-oxide aryl sulfones and sulfoxides |
Country Status (1)
| Country | Link |
|---|---|
| WO (1) | WO2008024692A1 (en) |
Cited By (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2008002849A3 (en) * | 2006-06-26 | 2008-12-04 | Neurogen Corp | Aryl sulfones |
| WO2009124734A3 (en) * | 2008-04-08 | 2010-04-01 | Grünenthal GmbH | Substituted sulfonamide derivatives |
| US20100317644A1 (en) * | 2006-09-29 | 2010-12-16 | Gruenenthal Gmbh | Substituted Sulfonamide Compounds |
| JP2012517457A (en) * | 2009-02-13 | 2012-08-02 | ベーリンガー インゲルハイム インターナショナル ゲゼルシャフト ミット ベシュレンクテル ハフツング | New preparation method |
| US8318769B2 (en) | 2008-04-08 | 2012-11-27 | Gruenethal Gmbh | Substituted sulfonamide compounds |
| WO2014029193A1 (en) | 2012-08-24 | 2014-02-27 | Sunshine Lake Pharma Co., Ltd. | Dihydropyrimidine compounds and their application in pharmaceuticals |
Citations (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2006113140A2 (en) * | 2005-04-15 | 2006-10-26 | Elan Pharmaceuticals, Inc. | Novel compounds useful for bradykinin b1 receptor antagonism |
-
2007
- 2007-08-20 WO PCT/US2007/076266 patent/WO2008024692A1/en not_active Ceased
Patent Citations (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2006113140A2 (en) * | 2005-04-15 | 2006-10-26 | Elan Pharmaceuticals, Inc. | Novel compounds useful for bradykinin b1 receptor antagonism |
Non-Patent Citations (1)
| Title |
|---|
| DATABASE CAPLUS [online] Database accession no. (2006:1124909) * |
Cited By (9)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2008002849A3 (en) * | 2006-06-26 | 2008-12-04 | Neurogen Corp | Aryl sulfones |
| US20100317644A1 (en) * | 2006-09-29 | 2010-12-16 | Gruenenthal Gmbh | Substituted Sulfonamide Compounds |
| US8435978B2 (en) * | 2006-09-29 | 2013-05-07 | Gruenenthal Gmbh | Substituted sulfonamide compounds |
| WO2009124734A3 (en) * | 2008-04-08 | 2010-04-01 | Grünenthal GmbH | Substituted sulfonamide derivatives |
| JP2011516514A (en) * | 2008-04-08 | 2011-05-26 | グリュネンタール・ゲゼルシャフト・ミト・ベシュレンクテル・ハフツング | Substituted sulfonamide derivatives |
| US8124624B2 (en) | 2008-04-08 | 2012-02-28 | Gruenenthal Gmbh | Substituted sulfonamide compounds |
| US8318769B2 (en) | 2008-04-08 | 2012-11-27 | Gruenethal Gmbh | Substituted sulfonamide compounds |
| JP2012517457A (en) * | 2009-02-13 | 2012-08-02 | ベーリンガー インゲルハイム インターナショナル ゲゼルシャフト ミット ベシュレンクテル ハフツング | New preparation method |
| WO2014029193A1 (en) | 2012-08-24 | 2014-02-27 | Sunshine Lake Pharma Co., Ltd. | Dihydropyrimidine compounds and their application in pharmaceuticals |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| WO2007140383A2 (en) | Spirocyclic sulfonamides and related compounds | |
| US8580812B2 (en) | Heteroaryl amide analogues as P2X7 antagonists | |
| AU2008286946B2 (en) | Heteroaryl amide analogues | |
| WO2007101007A2 (en) | Aryl sulfonyl heterocycles | |
| US20120190680A1 (en) | Heteroaryl Amide Analogues | |
| DK2178865T3 (en) | 5-membered heterocyclic amides and related compounds | |
| US20100216763A1 (en) | Heteroaryl Amide Derivatives | |
| WO2009108551A2 (en) | Heteroaryl amide analogues | |
| CA2660957C (en) | 2-phenoxy pyrimidinone analogues | |
| WO2008033739A2 (en) | Benzimidazole carboxamide derivatives | |
| US20080090845A1 (en) | Haloalkyl-substituted pyrimidinone derivatives | |
| CN111050765B (en) | Spirocyclic compounds and methods of making and using the same | |
| WO2008066664A2 (en) | Cis-cyclohexyl substituted pyrimidinone derivatives | |
| WO2008156607A1 (en) | Substituted pyrimidinones | |
| WO2008070014A2 (en) | Biaryl ketone-substituted piperidines | |
| WO2009121036A2 (en) | Substituted aryl pyrimidinone derivatives | |
| WO2008060568A2 (en) | Amide-substituted aryl piperidines | |
| WO2008002849A2 (en) | Aryl sulfones | |
| WO2009100403A1 (en) | Substituted aryl pyrimidinones |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 07814247 Country of ref document: EP Kind code of ref document: A1 |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| NENP | Non-entry into the national phase |
Ref country code: RU |
|
| 122 | Ep: pct application non-entry in european phase |
Ref document number: 07814247 Country of ref document: EP Kind code of ref document: A1 |