EP1701792B1 - Method for producing heteropolyacid catalyst - Google Patents
Method for producing heteropolyacid catalyst Download PDFInfo
- Publication number
- EP1701792B1 EP1701792B1 EP05704433.1A EP05704433A EP1701792B1 EP 1701792 B1 EP1701792 B1 EP 1701792B1 EP 05704433 A EP05704433 A EP 05704433A EP 1701792 B1 EP1701792 B1 EP 1701792B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- catalyst
- heteropolyacid
- pyridine
- heteropolyacid catalyst
- solution
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Expired - Lifetime
Links
Images

Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C51/00—Preparation of carboxylic acids or their salts, halides or anhydrides
- C07C51/16—Preparation of carboxylic acids or their salts, halides or anhydrides by oxidation
- C07C51/21—Preparation of carboxylic acids or their salts, halides or anhydrides by oxidation with molecular oxygen
- C07C51/25—Preparation of carboxylic acids or their salts, halides or anhydrides by oxidation with molecular oxygen of unsaturated compounds containing no six-membered aromatic ring
- C07C51/252—Preparation of carboxylic acids or their salts, halides or anhydrides by oxidation with molecular oxygen of unsaturated compounds containing no six-membered aromatic ring of propene, butenes, acrolein or methacrolein
-
- E—FIXED CONSTRUCTIONS
- E02—HYDRAULIC ENGINEERING; FOUNDATIONS; SOIL SHIFTING
- E02F—DREDGING; SOIL-SHIFTING
- E02F5/00—Dredgers or soil-shifting machines for special purposes
- E02F5/30—Auxiliary apparatus, e.g. for thawing, cracking, blowing-up, or other preparatory treatment of the soil
- E02F5/305—Arrangements for breaking-up hard ground
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J23/00—Catalysts comprising metals or metal oxides or hydroxides, not provided for in group B01J21/00
- B01J23/002—Mixed oxides other than spinels, e.g. perovskite
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J27/00—Catalysts comprising the elements or compounds of halogens, sulfur, selenium, tellurium, phosphorus or nitrogen; Catalysts comprising carbon compounds
- B01J27/14—Phosphorus; Compounds thereof
- B01J27/186—Phosphorus; Compounds thereof with arsenic, antimony, bismuth, vanadium, niobium, tantalum, polonium, chromium, molybdenum, tungsten, manganese, technetium or rhenium
- B01J27/188—Phosphorus; Compounds thereof with arsenic, antimony, bismuth, vanadium, niobium, tantalum, polonium, chromium, molybdenum, tungsten, manganese, technetium or rhenium with chromium, molybdenum, tungsten or polonium
- B01J27/19—Molybdenum
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J27/00—Catalysts comprising the elements or compounds of halogens, sulfur, selenium, tellurium, phosphorus or nitrogen; Catalysts comprising carbon compounds
- B01J27/14—Phosphorus; Compounds thereof
- B01J27/186—Phosphorus; Compounds thereof with arsenic, antimony, bismuth, vanadium, niobium, tantalum, polonium, chromium, molybdenum, tungsten, manganese, technetium or rhenium
- B01J27/195—Phosphorus; Compounds thereof with arsenic, antimony, bismuth, vanadium, niobium, tantalum, polonium, chromium, molybdenum, tungsten, manganese, technetium or rhenium with vanadium, niobium or tantalum
- B01J27/198—Vanadium
- B01J27/199—Vanadium with chromium, molybdenum, tungsten or polonium
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J2231/00—Catalytic reactions performed with catalysts classified in B01J31/00
- B01J2231/70—Oxidation reactions, e.g. epoxidation, (di)hydroxylation, dehydrogenation and analogues
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J2235/00—Indexing scheme associated with group B01J35/00, related to the analysis techniques used to determine the catalysts form or properties
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J2523/00—Constitutive chemical elements of heterogeneous catalysts
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J31/00—Catalysts comprising hydrides, coordination complexes or organic compounds
- B01J31/02—Catalysts comprising hydrides, coordination complexes or organic compounds containing organic compounds or metal hydrides
- B01J31/0234—Nitrogen-, phosphorus-, arsenic- or antimony-containing compounds
- B01J31/0235—Nitrogen containing compounds
- B01J31/0244—Nitrogen containing compounds with nitrogen contained as ring member in aromatic compounds or moieties, e.g. pyridine
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J35/00—Catalysts, in general, characterised by their form or physical properties
- B01J35/30—Catalysts, in general, characterised by their form or physical properties characterised by their physical properties
- B01J35/391—Physical properties of the active metal ingredient
- B01J35/393—Metal or metal oxide crystallite size
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J37/00—Processes, in general, for preparing catalysts; Processes, in general, for activation of catalysts
- B01J37/0009—Use of binding agents; Moulding; Pressing; Powdering; Granulating; Addition of materials ameliorating the mechanical properties of the product catalyst
-
- B—PERFORMING OPERATIONS; TRANSPORTING
- B01—PHYSICAL OR CHEMICAL PROCESSES OR APPARATUS IN GENERAL
- B01J—CHEMICAL OR PHYSICAL PROCESSES, e.g. CATALYSIS OR COLLOID CHEMISTRY; THEIR RELEVANT APPARATUS
- B01J37/00—Processes, in general, for preparing catalysts; Processes, in general, for activation of catalysts
- B01J37/02—Impregnation, coating or precipitation
- B01J37/0236—Drying, e.g. preparing a suspension, adding a soluble salt and drying
Definitions
- the present invention relates to a method for producing a phosphomolybdate-based heteropolyacid catalyst which is useful in vapor-phase oxidation of methacrolein, as represented by the following formula 1: PMo a A b B c C d D e E f O g (1) wherein A, B, C, D, E, a, b, c, d, e, f, and g are as defined in below.
- a heteropolyacid catalyst for partial oxidation of methacrolein to methacrylic acid by a vapor-phase oxidation process.
- a catalyst precursor is mainly prepared by coprecipitation with nitric acid or by vacuum drying in the absence of nitric acid.
- U.S. Patent No. 4,301,031 describes that a heteropolyacid catalyst with a composition of M0 12 P 0.1-3 M 0.1-3 Cu 0.1-2 V 0.1-2 X 0.01-2 Y a O b (M is K, Rb, or Cs; X is Ba, La, Ga, Al, Ag, Cd, Ti, Tl, Hg, Pb, or Zn; Y is Fe, Co, Ni, Sr, Mn, In, Ta, Ge, S, or Be) is useful for the oxidation of methacrolein.
- the heteropolyacid catalyst with the above-described composition is produced to a final catalyst by drying, forming into a predetermined shape, and calcination.
- pellets with a diameter of 5 mm and a length of 5 mm are generally formed.
- Decomposable ammonium or nitrate species are decomposed by the heat treatment to complete a catalyst with a desired structure and composition.
- the calcination is performed at a temperature of 300-500 °C under an oxygen or nitrogen atmosphere.
- a heteropolyacid catalyst production method varies according to the type of a metal-containing catalyst precursor.
- ammonium paramolybdate and ammonium metavanadate are mainly used.
- a catalyst production method disclosed in Example 12 of U.S. Patent No. 4,558,028 A follows the same manner as in Example 1 of U.S. Patent No. 6,333,293 B1 except that an appropriate amount of 85% phosphoric acid is added together with an appropriate amount of pyridine, and nitric acid is added before the addition of cesium nitrate and copper nitrate.
- U.S. Patent No. 6,458,740 B2 discloses a method for producing a heteropolyacid catalyst including adding pyridine and 85% phosphoric acid to a solution containing ammonium paramolybdate and ammonium metavanadate, adding nitric acid, cesium nitrate, and copper nitrate to the resultant solution to induce coprecipitation, followed by heating and drying.
- This patent describes that activity and selectivity of a catalyst are affected by the ratio of NH 4 /Mo 12 and NH 4 /NO 3 contained in catalyst precursors.
- JP 2000 202294 A provides a catalyst capable of producing methacrylic acid by vapor phase catalytic oxidation of methacrolein and produces methacrylic acid.
- Korean Patent No. KR 960003795 B1 describes that transition metal ions such as Cu ion can be efficiently precipitated using a mixture of pyridine with oxalic acid.
- Keggin structure is prerequisite for production of a heteropolyacid catalyst. Reaction of a molybdenum precursor, a vanadium precursor, and a phosphoric acid partially forms a precipitate but these components are partially present in their dissolved forms in water. At this time, when the pH of the solution is lowered using nitric acid or the like, crystals are created and precipitated. When nitric acid is dropwise added in the absence of a transition metal, an ammonium ion-containing heteropolyacid is formed. At this time, nitric acid can be precipitated together with another transition metal, which is called coprecipitation. Therefore, a Keggin anion structure is created and bonds with another transition metal as a cation to form a precipitate.
- transition metals may not form water-insoluble salts of them with heteropolyacid at low pH. Copper is the most representative metal. In this respect, as described above, even though coprecipitation is used for preparation of a heteropolyacid catalyst, copper ions remain in their dissolved forms without being precipitated and makes a blue color solution.
- a prepared catalyst slurry is dried in vacuum or heated above the boiling point of water so that metal ions are uniformly distributed in a catalyst simultaneously with drying of the slurry.
- the present inventors found a method for producing a heteropolyacid catalyst by preparing an ammonium ion-containing heteropolyacid using nitric acid followed by addition of a metal precursor, unlike a conventional catalyst production method by coaddition and coprecipitation of a transition metal and nitric acid.
- a transition metal carbonate instead of a common transition metal nitrate, is used as the metal precursor, dispersion of a metal component is enhanced, thereby increasing catalyst activity.
- the present inventors thus completed the present invention.
- the method of the present invention produces a heteropolyacid catalyst represented by formula 1.
- a heteropolyacid catalyst represented by the following formula 1 is obtained: PMo a A b B c C d D e E f O g (1) wherein A is V; B is an alkaline metal or an alkaline earth metal; C is Cu; D is Fe, Ce, Cr, Sn, Zn, Pd, or Rh; E is pyridine; a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f.
- the heteropolyacid catalyst of formula 1 obtained by the method of the present invention may have a composition of PMo 12 O 0.3 ⁇ 2 Cs 1 ⁇ 2 Cu 0 . 1 ⁇ 0 . 5 Fe 0.01 ⁇ 01 O x where x is a number satisfying the valence requirements.
- the method for producing the heteropolyacid catalyst as represented above by the formula 1, comprises:
- step (c) a transition metal nitrate may be further added.
- Particles formed on a surface of the heteropolyacid catalyst may have a particle size of 20-100 nm.
- the method of the present invention produces a heteropolyacid catalyst, wherein the heteropolyacid catalyst is represented by the following formula 2: PMo a V b Cs c Cu d Fe e pyridine f O g (2), wherein a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f.
- the heteropolyacid catalyst is represented by the following formula 2: PMo a V b Cs c Cu d Fe e pyridine f O g (2), wherein a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and
- Methacrylic acid which is a product of the oxidation of methacrolein by a heteropolyacid catalyst according to the present invention is used as a monomer of polymethylmethacrylate.
- composition of the heteropolyacid catalyst of the formula 1 is based on phosphomolybdate represented by the formula, H 3 PMo 12 O 40 .
- Phosphomolybdate has a Keggin structure in which one phosphate is surrounded by 12 octahedral molybdenum (Mo) oxides sharing edges.
- the molybdenum (Mo) may be partially or wholly substituted by oxide of the A element. That is, catalyst activity can be modified by an electron effect while maintaining the same catalyst structure as phosphomolybdate.
- the number of cations to be bonded is determined by the oxidation state or amount of an element to be substituted. For example, when a Mo +6 ion is substituted by a V +5 ion, the number of cations to be bonded is increased from 3 to 4.
- the hydrogen ions can be substituted by alkaline metals, alkaline earth metals, transition metals, ammonium ions, or pyridine ions.
- the elements B, C, and D of the formula 1 represent substituted cations.
- the substituted cations create a secondary or tertiary structure of a catalyst. Therefore, catalyst physical properties such as surface area, pore volume, and pore distribution can be adjusted, thereby increasing catalyst performance.
- a catalyst is dried at a temperature of 100- 150 °C.
- a dried catalyst is extruded on an extruder to prepare a catalyst with a predetermined shape.
- Extrusion is a very important process determining the physical strength of a catalyst.
- an appropriate amount of water, a glass fiber for reinforcement of catalyst strength, etc. may be used for extrusion.
- An extruded catalyst has a cylindrical shape with a diameter of about 5 mm and a length of about 5 mm.
- the cylindrical catalyst is calcined in a furnace at 350-500 °C in an air or nitrogen atmosphere for a predetermined time. During the calcination, nitrogen oxide contained in a catalyst precursor, such as ammonia and nitrate, is removed. Also, pyridine is partially removed and combined water is gradually removed.
- the most representative B element is a transition metal Cs
- the C element is Cu
- the most representative D element is a metal Fe.
- Cs and Cu are derived from carbonate precursors which are commercially available. Fe is derived from a nitrate precursor and the nitrate precursor is used in a small amount in catalyst production.
- the present inventors found that a catalyst with hexahedral crystal faces obtained by reaction of a previously prepared ammonium ion-containing heteropolyacid with a carbonate precursor is the most efficient catalyst.
- a catalyst with hexahedral crystal faces obtained by reaction of a previously prepared ammonium ion-containing heteropolyacid with a carbonate precursor is the most efficient catalyst.
- an ammonium ion-containing heteropolyacid is prepared and then reaction of the ammonium ion-containing heteropolyacid with a carbonate precursor is performed, a high efficiency catalyst can be produced.
- (NH4) 6 Mo 7 O 24 , (NH 4 )VO 3 , and H 3 PO 4 , and optionally pyridine are mixed to prepare an aqueous solution. That is, ammonium paramolybdate ((NH 4 ) 6 Mo 7 O 24 ) and ammonium metavanadate ((NH 4 )VO 3 ) are dissolved in distilled water and H 3 PO 4 is added thereto with stirring. Pyridine is selectively added to the resultant solution and stirred to prepare an aqueous solution.
- Nitric acid is added to the resultant solution of step (a) to prepare an ammonium ion-containing heteropolyacid. That is, HNO 3 is gradually dropwise added to the resultant solution of step (a) to make slurry. The slurry is stirred to prepare the ammonium ion-containing heteropolyacid.
- Copper carbonate is added to the ammonium ion-containing heteropolyacid obtained in step (b). That is, Cu(OH) 2 .
- CuCO 3 , Cs 2 CO 3 , and optionally Fe(NO 3 ) 3 are added to the ammonium ion-containing heteropolyacid obtained in step (b) at 20-60 °C and heated to 70 °C to increase catalyst dispersibility and activity.
- a catalyst-containing solution obtained in step (c) is dried and calcined in an air atmosphere. That is, the catalyst-containing solution obtained in step (c) is subjected to removal of excess water in a rotary evaporator and dried in an oven. Then, the resultant product is mixed with a glass fiber and formed into a pellet of 5 mm (diameter) x 5 mm (length), followed by calcination, to produce a catalyst with a particle size of 150-250 ⁇ m.
- the copper carbonate is added at a temperature of 20-60 °C. If the addition temperature of the copper carbonate is less than 20 °C, separate cooling is required and a production cost increases. On the other hand, if it exceeds 60 °C, decomposition of carbonate may occur.
- a solution of 60 ml of HNO 3 in 100 ml of H 2 O was gradually dropwise added to the resultant solution to make slurry.
- CuCO 3 and 12.55 g of Cs 2 CO 3 were added to the slurry at 40 °C with stirring and incubated at 70 °C .
- the resultant solution was subjected to removal of excess water in a rotary evaporator and dried in a 120°C oven.
- the dried product was mixed with 5 g of a glass fiber, formed into pellets of 5 mm (diameter) x 5 mm (length), calcined at 400 °C for 5 hours, and pulverized, to produce a catalyst with a particle size of 150-250 ⁇ m.
- a catalyst was prepared in the same manner as in Example 1 except that 1.04 g of Fe(NO 3 ) 3 was further added, in addition to 1.7 g (7.7 mmol) of Cu(OH) 2 . CuCO 3 and 12.55g (38.5 mmol) Cs 2 CO 3 .
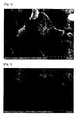
- the prepared catalyst had the composition of PM0 12 V 0.3 ⁇ 2 Cs 1v2 Cu 0 . 1v0 . 5 Fe 0.01 ⁇ 0 . 1 O x (x is a number satisfying the valence requirements) and a surface Scanning Electron Microscopy (SEM) image of the catalyst is shown in FIG. 1 .
- the SEM image shows that 11 scale bars are written in 200 nm, and thus, a distance between the bars is 20 nm. This is also applied to Figures illustrated in the following Examples and Comparative Examples.
- a catalyst was prepared in the same manner as in Example 2 except that pyridine was not used.
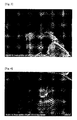
- a surface SEM image of the catalyst is shown in FIG. 2 .
- a catalyst was prepared in the same manner as in Example 1 except that 3.6g (15.4 mmol) of Cu(NO 3 ) 2 and 15.0g (77 mmol) of CsNO 3 were used instead of 1.7g (7.7 mmol) of Cu(OH) 2 .
- CuCO 3 and 12.55g (38.5 mmol) of Cs 2 CO 3 A surface SEM image of the catalyst is shown in FIG. 3 .
- a catalyst was prepared in the same manner as in Comparative Example 1 except that the transition metal precursors and nitric acid underwent coprecipitation like U.S. Patent No. 6,458,740B2 .
- a surface SEM image of the catalyst is shown in FIG. 4 .
- a commercially available catalyst was evaluated for catalyst performance and a surface SEM image of the catalyst is shown in FIG. 5 .
- catalysts with a particle size of 150-250 ⁇ m obtained by pulverization of calcined pellets were used to minimize the effect of the pressure change.
- Catalytic reaction was performed under the composition of 3.6 mol% of methacrolein (MACR), 10 mol% of H 2 O, 9.2 mol% of O 2 , and balance N 2 .
- a dose of a used catalyst was 1 g.
- the reaction temperature was 280-320 °C.
- a product was quantified by Gas Chromatography (GC).
- Methacrolein conversion rate % moles of reacted methacrolein / moles of supplied methacrolein x 100
- Selectivity % moles of produced methacrylic acid / moles of reacted methacrolein x 100
- the catalysts prepared in Examples 1-3 according to the present invention exhibited remarkably excellent conversion rate, selectivity, and yield, as compared with the catalysts of Comparative Examples 1-3 according to conventional technologies.
- particles of 20 to 100 nm in size were uniformly distributed on surfaces of the catalysts, relative to the catalysts of Comparative Examples 1-3. This shows that transition metal ions were uniformly dispersed.
- a heteropolyacid catalyst of the present invention is excellent in conversion rate, selectivity, and yield, relative to that produced by a conventional heteropolyacid catalyst production technology.
Landscapes
- Chemical & Material Sciences (AREA)
- Engineering & Computer Science (AREA)
- Organic Chemistry (AREA)
- Materials Engineering (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Oil, Petroleum & Natural Gas (AREA)
- Mechanical Engineering (AREA)
- Mining & Mineral Resources (AREA)
- Civil Engineering (AREA)
- General Engineering & Computer Science (AREA)
- Structural Engineering (AREA)
- Catalysts (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Description
- The present invention relates to a method for producing a phosphomolybdate-based heteropolyacid catalyst which is useful in vapor-phase oxidation of methacrolein, as represented by the following formula 1:
PMoaAbBcCdDeEfOg (1)
wherein A, B, C, D, E, a, b, c, d, e, f, and g are as defined in below. - Japanese companies such as Nippon Shokubai Chemical Ltd. (NSCL) hold a majority of patents about a heteropolyacid catalyst for partial oxidation of methacrolein to methacrylic acid by a vapor-phase oxidation process. In these patents, a catalyst precursor is mainly prepared by coprecipitation with nitric acid or by vacuum drying in the absence of nitric acid.
- Meanwhile,
U.S. Patent No. 4,301,031 describes that a heteropolyacid catalyst with a composition of M012P0.1-3M0.1-3Cu0.1-2V0.1-2X0.01-2YaOb (M is K, Rb, or Cs; X is Ba, La, Ga, Al, Ag, Cd, Ti, Tl, Hg, Pb, or Zn; Y is Fe, Co, Ni, Sr, Mn, In, Ta, Ge, S, or Be) is useful for the oxidation of methacrolein. The heteropolyacid catalyst with the above-described composition is produced to a final catalyst by drying, forming into a predetermined shape, and calcination. During the forming into a predetermined shape, pellets with a diameter of 5 mm and a length of 5 mm are generally formed. Decomposable ammonium or nitrate species are decomposed by the heat treatment to complete a catalyst with a desired structure and composition. The calcination is performed at a temperature of 300-500 °C under an oxygen or nitrogen atmosphere. -
U.S. Patent No. 4,621,155 describes that the preparation of a heteropolyacid catalyst in the presence of an N-containing material such as pyridine, piperidine, and piperazine can enhance formability and physical strength of the catalyst and reproducibility of the catalyst preparation. - A heteropolyacid catalyst production method varies according to the type of a metal-containing catalyst precursor. However, ammonium paramolybdate and ammonium metavanadate are mainly used.
- According to a catalyst production method disclosed in Example 1 of
U.S. Patent No. 6,333,293 B1 , ammonium paramolybdate and ammonium metavanadate are dissolved in heated water with stirring and an appropriate amount of 85% phosphoric acid is added thereto. Then, cesium nitrate and copper nitrate are added to the resultant solution followed by heating and drying to produce a catalyst. - A catalyst production method disclosed in Example 12 of
U.S. Patent No. 4,558,028 A follows the same manner as in Example 1 ofU.S. Patent No. 6,333,293 B1 except that an appropriate amount of 85% phosphoric acid is added together with an appropriate amount of pyridine, and nitric acid is added before the addition of cesium nitrate and copper nitrate. -
U.S. Patent No. 6,458,740 B2 discloses a method for producing a heteropolyacid catalyst including adding pyridine and 85% phosphoric acid to a solution containing ammonium paramolybdate and ammonium metavanadate, adding nitric acid, cesium nitrate, and copper nitrate to the resultant solution to induce coprecipitation, followed by heating and drying. This patent describes that activity and selectivity of a catalyst are affected by the ratio of NH4/Mo12 and NH4/NO3 contained in catalyst precursors. - Further,
JP 2000 202294 A - Meanwhile, Korean Patent No.
KR 960003795 B1 - In spite of these many inventions, since common heteropolyacid catalysts have low activity, there is an increased need to improve the conversion rate of methacrolein or selectivity to methacrylic acid. Furthermore, there is a problem in that productivity is lowered due to the low yield of a catalyst.
- In addition, formation of a Keggin structure is prerequisite for production of a heteropolyacid catalyst. Reaction of a molybdenum precursor, a vanadium precursor, and a phosphoric acid partially forms a precipitate but these components are partially present in their dissolved forms in water. At this time, when the pH of the solution is lowered using nitric acid or the like, crystals are created and precipitated. When nitric acid is dropwise added in the absence of a transition metal, an ammonium ion-containing heteropolyacid is formed. At this time, nitric acid can be precipitated together with another transition metal, which is called coprecipitation. Therefore, a Keggin anion structure is created and bonds with another transition metal as a cation to form a precipitate.
- Generally, it is known that coprecipitation leads to uniform precipitation of a transition metal. However, in a heteropolyacid production process, precipitation by nitric acid and transition metal salt formation are simultaneously performed, and thus, there is a high likelihood of non-uniform precipitation.
- Some transition metals may not form water-insoluble salts of them with heteropolyacid at low pH. Copper is the most representative metal. In this respect, as described above, even though coprecipitation is used for preparation of a heteropolyacid catalyst, copper ions remain in their dissolved forms without being precipitated and makes a blue color solution.
- To solve these problems, a prepared catalyst slurry is dried in vacuum or heated above the boiling point of water so that metal ions are uniformly distributed in a catalyst simultaneously with drying of the slurry.
- While searching for solutions to these problems, the present inventors found a method for producing a heteropolyacid catalyst by preparing an ammonium ion-containing heteropolyacid using nitric acid followed by addition of a metal precursor, unlike a conventional catalyst production method by coaddition and coprecipitation of a transition metal and nitric acid. At this time, when a transition metal carbonate, instead of a common transition metal nitrate, is used as the metal precursor, dispersion of a metal component is enhanced, thereby increasing catalyst activity. The present inventors thus completed the present invention.
- Therefore, the method of the present invention produces a heteropolyacid catalyst represented by formula 1.
- The above and other objects of the present invention can be accomplished by embodiments of the present invention as will be described hereinafter.
- According to the method of the present invention, a heteropolyacid catalyst represented by the following formula 1 is obtained:
PMoaAbBcCdDeEfOg (1)
wherein A is V; B is an alkaline metal or an alkaline earth metal; C is Cu; D is Fe, Ce, Cr, Sn, Zn, Pd, or Rh; E is pyridine; a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f. - The heteropolyacid catalyst of formula 1 obtained by the method of the present invention may have a composition of PMo12O0.3~2Cs1~2Cu0.1~0.5Fe0.01~01Ox where x is a number satisfying the valence requirements.
- The method for producing the heteropolyacid catalyst as represented above by the formula 1, comprises:
- (a) mixing (NH4)6Mo7O24, (NH4)VO3, and H3PO4, and optionally pyridine, to prepare an aqueous solution;
- (b) adding nitric acid to the solution of step (a) to prepare an ammonium ion-containing heteropolyacid;
- (c) adding copper carbonate at a temperature of 20-60 °C to a solution obtained in step (b); and
- (d) drying and calcining a catalyst-containing solution obtained in step (c) in an air atmosphere.
- In step (c), a transition metal nitrate may be further added.
- Particles formed on a surface of the heteropolyacid catalyst may have a particle size of 20-100 nm.
- In a further embodiment, the method of the present invention produces a heteropolyacid catalyst, wherein the heteropolyacid catalyst is represented by the following formula 2:
PMoaVbCscCudFeepyridinefOg (2),
wherein a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f. - Hereinafter, the present invention will be described in detail.
- Methacrylic acid which is a product of the oxidation of methacrolein by a heteropolyacid catalyst according to the present invention is used as a monomer of polymethylmethacrylate.
- The composition of the heteropolyacid catalyst of the formula 1 is based on phosphomolybdate represented by the formula, H3PMo12O40. Phosphomolybdate has a Keggin structure in which one phosphate is surrounded by 12 octahedral molybdenum (Mo) oxides sharing edges.
- The molybdenum (Mo) may be partially or wholly substituted by oxide of the A element. That is, catalyst activity can be modified by an electron effect while maintaining the same catalyst structure as phosphomolybdate. The number of cations to be bonded is determined by the oxidation state or amount of an element to be substituted. For example, when a Mo+6 ion is substituted by a V+5 ion, the number of cations to be bonded is increased from 3 to 4. The hydrogen ions can be substituted by alkaline metals, alkaline earth metals, transition metals, ammonium ions, or pyridine ions. The elements B, C, and D of the formula 1 represent substituted cations.
- The substituted cations create a secondary or tertiary structure of a catalyst. Therefore, catalyst physical properties such as surface area, pore volume, and pore distribution can be adjusted, thereby increasing catalyst performance.
- Generally, a catalyst is dried at a temperature of 100- 150 °C.
- A dried catalyst is extruded on an extruder to prepare a catalyst with a predetermined shape. Extrusion is a very important process determining the physical strength of a catalyst. In this respect, an appropriate amount of water, a glass fiber for reinforcement of catalyst strength, etc. may be used for extrusion. An extruded catalyst has a cylindrical shape with a diameter of about 5 mm and a length of about 5 mm. The cylindrical catalyst is calcined in a furnace at 350-500 °C in an air or nitrogen atmosphere for a predetermined time. During the calcination, nitrogen oxide contained in a catalyst precursor, such as ammonia and nitrate, is removed. Also, pyridine is partially removed and combined water is gradually removed.
- In the heteropolyacid catalyst of the formula 1, the most representative B element is a transition metal Cs, the C element is Cu, and the most representative D element is a metal Fe. Cs and Cu are derived from carbonate precursors which are commercially available. Fe is derived from a nitrate precursor and the nitrate precursor is used in a small amount in catalyst production.
- The present inventors found that a catalyst with hexahedral crystal faces obtained by reaction of a previously prepared ammonium ion-containing heteropolyacid with a carbonate precursor is the most efficient catalyst. In this respect, when an ammonium ion-containing heteropolyacid is prepared and then reaction of the ammonium ion-containing heteropolyacid with a carbonate precursor is performed, a high efficiency catalyst can be produced.
- The method for producing a heteropolyacid catalyst according to the present invention will now be described schematically.
- (NH4)6Mo7O24, (NH4)VO3, and H3PO4, and optionally pyridine are mixed to prepare an aqueous solution. That is, ammonium paramolybdate ((NH4)6Mo7O24) and ammonium metavanadate ((NH4)VO3) are dissolved in distilled water and H3PO4 is added thereto with stirring. Pyridine is selectively added to the resultant solution and stirred to prepare an aqueous solution.
- Nitric acid is added to the resultant solution of step (a) to prepare an ammonium ion-containing heteropolyacid. That is, HNO3 is gradually dropwise added to the resultant solution of step (a) to make slurry. The slurry is stirred to prepare the ammonium ion-containing heteropolyacid.
- Copper carbonate is added to the ammonium ion-containing heteropolyacid obtained in step (b). That is, Cu(OH)2 . CuCO3, Cs2CO3, and optionally Fe(NO3)3 are added to the ammonium ion-containing heteropolyacid obtained in step (b) at 20-60 °C and heated to 70 °C to increase catalyst dispersibility and activity.
- A catalyst-containing solution obtained in step (c) is dried and calcined in an air atmosphere. That is, the catalyst-containing solution obtained in step (c) is subjected to removal of excess water in a rotary evaporator and dried in an oven. Then, the resultant product is mixed with a glass fiber and formed into a pellet of 5 mm (diameter) x 5 mm (length), followed by calcination, to produce a catalyst with a particle size of 150-250 µm.
- The copper carbonate is added at a temperature of 20-60 °C. If the addition temperature of the copper carbonate is less than 20 °C, separate cooling is required and a production cost increases. On the other hand, if it exceeds 60 °C, decomposition of carbonate may occur.
- Hereinafter, the present invention will be described more specifically by Examples. However, the following Examples are provided only for illustrations and thus the present invention is not limited to or by them.
-
-
FIG. 1 is a surface Scanning Electron Microscopy (SEM) image of a catalyst prepared in Example 2 according to the method of the present invention. -
FIG. 2 is a surface SEM image of a catalyst prepared in Example 3 according to the method of the present invention. -
FIG. 3 is a surface SEM image of a catalyst prepared in Comparative Example 1. -
FIG. 4 is a surface SEM image of a catalyst prepared in Comparative Example 2. -
FIG. 5 is a surface SEM image of a common catalyst of Comparative Example 3. - 100 g of ammonium paramolybdate ((NH4)6Mo7O24) and 3.0 g of ammonium metavanadate ((NH4)VO3) were dissolved in 440 ml of 80 °C distilled water. 23 g of pyridine and 6.4 g of 85% H3PO4 were added thereto with stirring.
- A solution of 60 ml of HNO3 in 100 ml of H2O was gradually dropwise added to the resultant solution to make slurry. 1.7 g of Cu(OH)2 . CuCO3 and 12.55 g of Cs2CO3 were added to the slurry at 40 °C with stirring and incubated at 70 °C . Then, the resultant solution was subjected to removal of excess water in a rotary evaporator and dried in a 120°C oven. The dried product was mixed with 5 g of a glass fiber, formed into pellets of 5 mm (diameter) x 5 mm (length), calcined at 400 °C for 5 hours, and pulverized, to produce a catalyst with a particle size of 150-250 µm.
- A catalyst was prepared in the same manner as in Example 1 except that 1.04 g of Fe(NO3)3 was further added, in addition to 1.7 g (7.7 mmol) of Cu(OH)2 . CuCO3 and 12.55g (38.5 mmol) Cs2CO3.
- The prepared catalyst had the composition of PM012V0.3~2Cs1v2Cu0.1v0.5Fe0.01~0.1Ox (x is a number satisfying the valence requirements) and a surface Scanning Electron Microscopy (SEM) image of the catalyst is shown in
FIG. 1 . The SEM image shows that 11 scale bars are written in 200 nm, and thus, a distance between the bars is 20 nm. This is also applied to Figures illustrated in the following Examples and Comparative Examples. - A catalyst was prepared in the same manner as in Example 2 except that pyridine was not used. A surface SEM image of the catalyst is shown in
FIG. 2 . - A catalyst was prepared in the same manner as in Example 1 except that 3.6g (15.4 mmol) of Cu(NO3)2 and 15.0g (77 mmol) of CsNO3 were used instead of 1.7g (7.7 mmol) of Cu(OH)2 . CuCO3 and 12.55g (38.5 mmol) of Cs2CO3. A surface SEM image of the catalyst is shown in
FIG. 3 . - A catalyst was prepared in the same manner as in Comparative Example 1 except that the transition metal precursors and nitric acid underwent coprecipitation like
U.S. Patent No. 6,458,740B2 . A surface SEM image of the catalyst is shown inFIG. 4 . - A commercially available catalyst was evaluated for catalyst performance and a surface SEM image of the catalyst is shown in
FIG. 5 . - Catalytic reaction results for the catalysts of Examples 1-3 and Comparative Examples 1-3 are presented in Table 1 below.
- For catalyst activity evaluation, catalysts with a particle size of 150-250 µm obtained by pulverization of calcined pellets were used to minimize the effect of the pressure change.
- Catalytic reaction was performed under the composition of 3.6 mol% of methacrolein (MACR), 10 mol% of H2O, 9.2 mol% of O2, and balance N2. A dose of a used catalyst was 1 g. The reaction temperature was 280-320 °C. A product was quantified by Gas Chromatography (GC).
- Methacrolein conversion rate, and methacrylic acid yield and selectivity were respectively calculated by Equations 1-3 below:


Table 1 Example Reaction temperature (°C) Conversion rate (%) Selectivity (%) Yield (%) Example 1 280 34.01 80.30 27.31 300 45.24 81.54 36.89 Example 280 36.19 80.16 29.01 300 49.58 80.99 40.16 Example 3 280 34.32 73.67 25.28 Comparative Example 1 280 21.07 66.71 14.06 300 31.98 74.17 23.72 Comparative Example 2 280 15.49 51.41 8.45 300 28.34 67.29 19.07 Comparative Example 3 280 30.13 76.19 22.95 - As can be seen from Table 1, the catalysts prepared in Examples 1-3 according to the present invention exhibited remarkably excellent conversion rate, selectivity, and yield, as compared with the catalysts of Comparative Examples 1-3 according to conventional technologies. In connection with the catalysts shown in
FIGS. 1 and 2 according to the present invention, particles of 20 to 100 nm in size were uniformly distributed on surfaces of the catalysts, relative to the catalysts of Comparative Examples 1-3. This shows that transition metal ions were uniformly dispersed. - As apparent from the above description, a heteropolyacid catalyst of the present invention is excellent in conversion rate, selectivity, and yield, relative to that produced by a conventional heteropolyacid catalyst production technology.
Claims (3)
- A method for producing a heteropolyacid catalyst represented by the
following formula 1:
PMoaAbBcCdDeEfOg (1),
wherein A is V; B is an alkaline metal or an alkaline earth metal; C is Cu; D is Fe, Ce, Cr, Sn, Zn, Pd, or Rh; E is pyridine; a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f,
the method comprising:(a) mixing (NH4)6Mo7O24, (NH4)VO3, and H3PO4, and optionally pyridine, to prepare an aqueous solution;(b) adding nitric acid to the solution of step (a) to prepare an ammonium ion-containing heteropolyacid;(c) adding copper carbonate at a temperature of 20-60 °C to a solution obtained in step (b); and(d) drying and calcining a catalyst-containing solution obtained in step (c) in an air atmosphere. - The method of claim 1, wherein particles formed on a surface of the heteropolyacid catalyst have a particle size of 20-100 nm.
- The method of claim 1, wherein the heteropolyacid catalyst is represented by the following formula 2:
PMoaVbCscCudFeepyridinefOg (2),
wherein a is 5-12; b is 0.01-5; c is 0.01-3; d is 0.01-3; e is 0-0.3; f is 0-10; and g is a number satisfying the valence requirements of a, b, c, d, e, and f.
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| KR1020040001574A KR100557640B1 (en) | 2004-01-09 | 2004-01-09 | Novel heteropolyacid catalyst and preparation method thereof |
| PCT/KR2005/000008 WO2005065822A1 (en) | 2004-01-09 | 2005-01-04 | Novel heteropolyacid catalyst and method for producing the same |
Publications (3)
| Publication Number | Publication Date |
|---|---|
| EP1701792A1 EP1701792A1 (en) | 2006-09-20 |
| EP1701792A4 EP1701792A4 (en) | 2010-09-22 |
| EP1701792B1 true EP1701792B1 (en) | 2014-12-24 |
Family
ID=36383764
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP05704433.1A Expired - Lifetime EP1701792B1 (en) | 2004-01-09 | 2005-01-04 | Method for producing heteropolyacid catalyst |
Country Status (7)
| Country | Link |
|---|---|
| US (1) | US7094727B2 (en) |
| EP (1) | EP1701792B1 (en) |
| JP (1) | JP2006513033A (en) |
| KR (1) | KR100557640B1 (en) |
| CN (1) | CN100360230C (en) |
| TW (1) | TWI292339B (en) |
| WO (1) | WO2005065822A1 (en) |
Families Citing this family (8)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN1590422A (en) * | 1999-12-23 | 2005-03-09 | 巴塞尔聚烯烃有限公司 | Transition metal compounds, ligand systems, catalyst systems and their use in the polymerization and copolymerization of olefins |
| EP1847555A1 (en) | 2006-04-18 | 2007-10-24 | Borealis Technology Oy | Multi-branched Polypropylene |
| KR101009893B1 (en) | 2007-04-06 | 2011-01-20 | 주식회사 엘지화학 | Novel Process for Preparing High Purity Heteropoly Acid |
| KR100954049B1 (en) | 2007-06-13 | 2010-04-20 | 주식회사 엘지화학 | Method for preparing heteropolyacid catalyst |
| CN106957313B (en) * | 2017-04-08 | 2018-01-02 | 山东本源晶体科技有限公司 | Preparation method and application of a heteropolyacid crystal |
| MY192580A (en) * | 2017-07-10 | 2022-08-29 | Mitsubishi Chem Corp | Method for producing catalyst, method for producing unsaturated aldehyde and unsaturated calboxylic acid, and method for producing unsaturated calboxylic acid ester |
| CN117324013B (en) * | 2022-06-24 | 2025-11-25 | 中国石油化工股份有限公司 | A catalyst suitable for the production of methacrylic acid, its preparation method and application |
| CN116196950B (en) * | 2023-02-16 | 2024-05-24 | 北华大学 | A synthesis method of a bimetallic doped heteropolyacid catalyst and its application in lignin conversion |
Family Cites Families (22)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US4056487A (en) * | 1975-10-02 | 1977-11-01 | Petro-Tex Chemical Corporation | Vanadium phosphorus oxygen oxidation catalysts useful for preparing anhydrides from alkanes |
| US4419270A (en) | 1980-06-26 | 1983-12-06 | Nippon Shokubai Kagaku Kogyo Co. Ltd. | Oxidation catalyst |
| US4301031A (en) | 1980-08-05 | 1981-11-17 | The Standard Oil Company | Methacrolein oxidation catalysts |
| JPS59115750A (en) | 1982-12-22 | 1984-07-04 | Nippon Shokubai Kagaku Kogyo Co Ltd | Catalyst for synthesis of methacrylic acid |
| GB2138694B (en) * | 1983-03-24 | 1986-12-10 | Nippon Catalytic Chem Ind | Heteropolyacid-type catalyst composition containing whiskers |
| JPH085820B2 (en) * | 1988-04-05 | 1996-01-24 | 旭化成工業株式会社 | Method for producing methacrylic acid and / or methacrolein |
| JP3101821B2 (en) * | 1990-02-15 | 2000-10-23 | 三菱レイヨン株式会社 | Preparation of catalyst for methacrylic acid production |
| JPH047037A (en) * | 1990-04-23 | 1992-01-10 | Mitsubishi Rayon Co Ltd | Preparation of catalyst for manufacturing methacrylic acid |
| FR2680702B1 (en) * | 1991-09-03 | 1994-11-04 | Atochem | NOVEL CATALYST SYSTEM AND ITS APPLICATION TO THE OXIDE OF HYDROGENATION OF SATURATED CARBOXYLIC ACIDS AND THE OXIDATION OF ALDEHYDES TO ACIDS. |
| KR960003795B1 (en) | 1993-04-13 | 1996-03-22 | 재단법인 한국화학연구소 | Method for preparing heteropolyacid catalyst |
| KR100210642B1 (en) * | 1994-05-31 | 1999-07-15 | 겐지 아이다 | Methacrylic acid production catalyst and method for producing methacrylic acid using this catalyst |
| JP2944463B2 (en) * | 1994-05-31 | 1999-09-06 | 株式会社日本触媒 | Catalyst for producing methacrylic acid and method for producing methacrylic acid using the catalyst |
| KR0142013B1 (en) | 1994-07-04 | 1998-06-01 | 김상응 | Method of preparing synthetic adsorbent |
| KR100204729B1 (en) * | 1997-03-17 | 1999-06-15 | 성재갑 | Method for producing a catalyst for acrolein partial oxidation |
| JP3710944B2 (en) * | 1999-01-19 | 2005-10-26 | 三菱レイヨン株式会社 | Catalyst for producing methacrylic acid, method for producing the same, and method for producing methacrylic acid |
| JP3734403B2 (en) * | 1999-04-27 | 2006-01-11 | 住友化学株式会社 | Catalyst for producing methacrylic acid and method for producing methacrylic acid |
| JP3690939B2 (en) * | 1999-06-07 | 2005-08-31 | 三菱レイヨン株式会社 | Catalyst for synthesizing methacrylic acid and method for producing methacrylic acid |
| JP3702710B2 (en) | 1999-06-15 | 2005-10-05 | 住友化学株式会社 | Catalyst for producing methacrylic acid and method for producing methacrylic acid |
| JP3767309B2 (en) * | 2000-03-07 | 2006-04-19 | 住友化学株式会社 | Method for producing heteropolyacid catalyst and method for producing methacrylic acid |
| JP3763246B2 (en) * | 2000-04-06 | 2006-04-05 | 住友化学株式会社 | Method for regenerating heteropolyacid catalyst and method for producing methacrylic acid |
| JP4671320B2 (en) * | 2000-09-21 | 2011-04-13 | 日本化薬株式会社 | Production of coated catalyst for methacrylic acid production |
| MXPA03002374A (en) * | 2000-09-21 | 2003-06-30 | Nippon Kayaku Kabushiki Kaisha | Catalyst for methacrylic acid production, coated catalyst, and process for producing the same. |
-
2004
- 2004-01-09 KR KR1020040001574A patent/KR100557640B1/en not_active Expired - Lifetime
- 2004-12-30 US US11/024,809 patent/US7094727B2/en not_active Expired - Lifetime
- 2004-12-31 TW TW093141798A patent/TWI292339B/en not_active IP Right Cessation
-
2005
- 2005-01-04 WO PCT/KR2005/000008 patent/WO2005065822A1/en not_active Ceased
- 2005-01-04 CN CNB2005800000265A patent/CN100360230C/en not_active Expired - Fee Related
- 2005-01-04 EP EP05704433.1A patent/EP1701792B1/en not_active Expired - Lifetime
- 2005-01-04 JP JP2005518334A patent/JP2006513033A/en active Pending
Also Published As
| Publication number | Publication date |
|---|---|
| EP1701792A1 (en) | 2006-09-20 |
| CN1798607A (en) | 2006-07-05 |
| KR100557640B1 (en) | 2006-03-10 |
| WO2005065822A1 (en) | 2005-07-21 |
| TWI292339B (en) | 2008-01-11 |
| US20050153831A1 (en) | 2005-07-14 |
| EP1701792A4 (en) | 2010-09-22 |
| US7094727B2 (en) | 2006-08-22 |
| JP2006513033A (en) | 2006-04-20 |
| TW200523021A (en) | 2005-07-16 |
| KR20050073224A (en) | 2005-07-13 |
| CN100360230C (en) | 2008-01-09 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP2075064B1 (en) | Fluidized-bed catalyst for producing acrylonitrile and process for producing acrylonitrile | |
| DE69101685T2 (en) | Iron-antimony-molybdenum catalytic composition and process for producing the same. | |
| KR101772247B1 (en) | Improved mixed metal oxide ammoxidation catalysts | |
| EP0460932B1 (en) | Method for preparing acrolein or methacrolein | |
| EP2100663B1 (en) | Catalyst for synthesizing acrylonitrile and process for producing acrylonitrile | |
| EP2098288B1 (en) | Catalyst for producing acrylonitrile and process for producing acrlyonitrile | |
| EP0005067A2 (en) | Technique for forming multi-component oxide complex catalysts | |
| JP4954750B2 (en) | Method for producing molybdenum, bismuth, iron, silica-containing composite oxide catalyst | |
| JP3371112B2 (en) | Iron / antimony-containing metal oxide catalyst composition and method for producing the same | |
| US8053386B2 (en) | Heteropoly acid catalyst and method of preparing the same | |
| EP0010429B1 (en) | Method for preparing an oxidation catalyst containing a molybdate anion | |
| WO2005079980A1 (en) | Catalyst for partial oxidation and preparation method thereof | |
| EP1503856B1 (en) | Method for preparing a catalyst for partial oxidation of propylene | |
| US7094727B2 (en) | Heteropolyacid catalyst and method for producing the same | |
| EP1613432B1 (en) | Method for preparing catalysts for partial oxidation of propylene and iso-butylene | |
| US7449426B2 (en) | Catalyst composition without antimony or molybdenum for ammoxidation of alkanes, a process of making and a process of using thereof | |
| KR100865654B1 (en) | Methacrylic acid production catalyst and methacrylic acid production method | |
| KR20140133862A (en) | Molybdenum mixed metal oxide catalysts for the production of unsaturated aldehydes from olefins | |
| JP3347263B2 (en) | Preparation of catalysts for the production of unsaturated aldehydes and unsaturated carboxylic acids | |
| KR100760461B1 (en) | Process for preparing heteropolyacid catalyst | |
| JP3720626B2 (en) | Method for preparing molybdenum-bismuth-tellurium-containing composite oxide catalyst | |
| KR100878688B1 (en) | Heat treatment method of heteropolyacid catalyst |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| 17P | Request for examination filed |
Effective date: 20050623 |
|
| AK | Designated contracting states |
Kind code of ref document: A1 Designated state(s): AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IS IT LI LT LU MC NL PL PT RO SE SI SK TR |
|
| DAX | Request for extension of the european patent (deleted) | ||
| A4 | Supplementary search report drawn up and despatched |
Effective date: 20100823 |
|
| RIC1 | Information provided on ipc code assigned before grant |
Ipc: B01J 37/00 20060101ALI20100813BHEP Ipc: B01J 23/89 20060101AFI20050725BHEP |
|
| 17Q | First examination report despatched |
Effective date: 20101108 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R079 Ref document number: 602005045504 Country of ref document: DE Free format text: PREVIOUS MAIN CLASS: B01J0023890000 Ipc: B01J0027190000 |
|
| GRAP | Despatch of communication of intention to grant a patent |
Free format text: ORIGINAL CODE: EPIDOSNIGR1 |
|
| RIC1 | Information provided on ipc code assigned before grant |
Ipc: B01J 27/19 20060101AFI20140826BHEP |
|
| INTG | Intention to grant announced |
Effective date: 20140911 |
|
| GRAS | Grant fee paid |
Free format text: ORIGINAL CODE: EPIDOSNIGR3 |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| AK | Designated contracting states |
Kind code of ref document: B1 Designated state(s): AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IS IT LI LT LU MC NL PL PT RO SE SI SK TR |
|
| REG | Reference to a national code |
Ref country code: GB Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: EP |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: AT Ref legal event code: REF Ref document number: 702862 Country of ref document: AT Kind code of ref document: T Effective date: 20150115 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R096 Ref document number: 602005045504 Country of ref document: DE Effective date: 20150219 |
|
| REG | Reference to a national code |
Ref country code: NL Ref legal event code: VDEP Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: FI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: LT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| REG | Reference to a national code |
Ref country code: LT Ref legal event code: MG4D |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20150325 Ref country code: SE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| REG | Reference to a national code |
Ref country code: AT Ref legal event code: MK05 Ref document number: 702862 Country of ref document: AT Kind code of ref document: T Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: NL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: RO Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: SK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: CZ Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: EE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: ES Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20150424 Ref country code: AT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: LU Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20150104 Ref country code: PL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R097 Ref document number: 602005045504 Country of ref document: DE |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MC Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LI Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20150131 Ref country code: CH Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20150131 Ref country code: DK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| REG | Reference to a national code |
Ref country code: FR Ref legal event code: ST Effective date: 20150930 |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: MM4A |
|
| GBPC | Gb: european patent ceased through non-payment of renewal fee |
Effective date: 20150324 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: FR Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20150224 |
|
| 26N | No opposition filed |
Effective date: 20150925 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GB Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20150324 Ref country code: IE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20150104 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BG Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 Ref country code: HU Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT; INVALID AB INITIO Effective date: 20050104 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: CY Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: PT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20150424 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: TR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20141224 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 20221220 Year of fee payment: 19 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R119 Ref document number: 602005045504 Country of ref document: DE |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240801 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20240801 |