WO2017134684A2 - An improved process for the preparation of ibrutinib - Google Patents
An improved process for the preparation of ibrutinib Download PDFInfo
- Publication number
- WO2017134684A2 WO2017134684A2 PCT/IN2017/050045 IN2017050045W WO2017134684A2 WO 2017134684 A2 WO2017134684 A2 WO 2017134684A2 IN 2017050045 W IN2017050045 W IN 2017050045W WO 2017134684 A2 WO2017134684 A2 WO 2017134684A2
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- ibrutinib
- pyrazolo
- phenoxyphenyl
- pyrimidine
- boc
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
- C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
- C07D487/04—Ortho-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D473/00—Heterocyclic compounds containing purine ring systems
- C07D473/02—Heterocyclic compounds containing purine ring systems with oxygen, sulphur, or nitrogen atoms directly attached in positions 2 and 6
- C07D473/04—Heterocyclic compounds containing purine ring systems with oxygen, sulphur, or nitrogen atoms directly attached in positions 2 and 6 two oxygen atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07B—GENERAL METHODS OF ORGANIC CHEMISTRY; APPARATUS THEREFOR
- C07B2200/00—Indexing scheme relating to specific properties of organic compounds
- C07B2200/13—Crystalline forms, e.g. polymorphs
Definitions
- the present invention relates to an improved process for the preparation of Ibrutinib with high purity and high yields.
- the present Invention also relates to a process for the preparation of Crystalline form A and form C of Ibrutinib.
- Ibrutinib is a kinase inhibitor indicated for the treatment of mantle cell lymphoma, chronic lymphocytic leukaemia and Waldenstrom's macroglobulinemia.
- Ibrutinib is chemically is known as l-[(3R)-3-[4-amino-3-(4-phenoxyphenyl)-lH- pyrazolo[3,4-d]pyrimidin-l-yl]-l-piperidinyl]-2-propen-l-one and structurally represented as below.
- Ibrutinib is first disclosed in US 7514444 and marketed as IMBRUVICA ® .
- US '444 patent is disclosed the Ibrutinib process, wherein 4-amino-3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine is reacted with tert-Butyl 3- hydroxypiperidine-l-carboxylate and diisopropyl diazodicarboxylate to obtain intermediate under mitsunobu reaction conditions by converting the hydroxy moiety of the tert-Butyl 3-hydroxypiperidine-l-carboxylate to a better leaving group, thereby allowing a substitution reaction. Further it is converted to ibrutinib by deprotection of Boc and acylation with acryl chloride.
- US9296753 patent has disclosed the crystalline forms A, B, C, D, E and F and their process for the preparation.
- the present inventors of the present invention has developed an improved process for the preparation of Ibrutinib with high yield and high purity.
- the present process is cost effective and feasible in large scale production also.
- the present process avoids the mitsunobu reagent conditions also.
- One aspect of the present invention is to provide a process for the preparation of ibrutinib comprising the steps of: a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
- Another aspect of the present Invention is to provide a process for the preparation of Ibrutinib Crystalline form A comprising the steps of:
- Yet another aspect of the present Invention is to provide a process for preparation of Ibrutinib Crystalline form C comprising the steps of: a) reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
- Yet another aspect of the present invention is to provide a purification of ibrutinib comprising the steps of:
- Yet another aspect of the present invention is to provide a process for the preparation of Crystalline form A of Ibrutinib comprising the steps of:
- Yet another aspect of the present invention is to provide a process for preparation of Crystalline form C of Ibrutinib comprising the steps of:
- Yet another aspect of the present invention is to provide a process for purification of ibrutinib comprising the steps of: a) dissolving ibrutinib in methylene chloride and toluene, and
- the present invention is related to an improved process for the preparation of Ibrutinib, wherein reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine and treating (S)-l-boc-3- methylsulfonyloxy piperidine with 3-(4-phenoxyphenyl)-lH-pyrazolo[3,4- d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine- 4-amine hydrochloride and converted to ibrutinib reacting with acryloyl chloride. Further ibrutinib is converted to crystalline form A and crystalline form C.
- triethylamine is added to the reaction mixture of (S)-l-Boc-3-hydroxypiperidin and methanesulfonyl chloride in methylenechloride at -10 to 10°C.
- the reaction mass is allowed to ambient temperature and stirred for 30 min and washed with water
- the organic layer is concentrated, stirred with hexane, filtered the solid and dried to yield (S)-l-boc-3- methylsulfonyloxy piperidine.
- the concentrated mass is stirred with methanol and ethyl acetate, filtered the solid and dried.
- the resulting dried product is dissolved in DM water and added to sodium carbonate solution and Ethyl acetate mixture.
- the product is filtered, washed with DM water and then the wet product is dissolved in Con.HCl and methanol mixture.
- the resulting reaction mass is concentrated, stirred with methanol and ethyl acetate
- the product is filtered, washed and dried to to yield (R)-3-(4-phenoxyphenyl)-l- (piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride.
- reaction mass is stirred for 10 min at -80 to -90°C and washed with aqueous citric acid solution followed by aqueous sodium bicarbonate solution and water.
- Inorganic bases is selected from cesium carbonate, potassium carbonate, sodium carbonate, sodium bicarbonate, sodium phosphate, potassium phosphate, potassium phosphate, sodium acetate, potassium acetate, preferably potassium carbonate and cesium carbonate.
- base employed in step c) is selected from diisopropylethylamine and trimethylamine, preferably diisopropylethylamine.
- Another embodiment of the present invention is to provide a purification of ibrutinib comprising the steps of: a) dissolving ibrutinib in non-polar solvent or polar aprotic solvent or mixture thereof and
- non-polar solvent is selected from pentane, hexane, toluene and cyclophexane, preferably hexane.
- polar aprotic solvent is selected from tetrahydrofuran, ethyl acetate and dimethyl formamide, preferably ethylacetate.
- polar aprotic solvent is selected from methylene chloride, ethyl acetate, acetone preferably ethyl acetate.
- polar protic solvent is selected from methanol, ethanol, n-propanol preferably methanol.
- Example-1 PREPARATION OF (S)-l-boc-3-METHYLSULFONYLOXY PIPERIDINE
- Triethylamine (120.7 g) was added to the reaction mixture of (S)-l-Boc-3- hydroxypiperidin (200 g) and methanesulfonyl chloride (125.3g) in methylenechloride (2000 ml) at 0°C.
- the reaction mass was allowed to ambient temperature and stirred for 30 min and washed with water (2 x 2 1).
- the organic layer was concentrated, stirred with hexane (1.4 1), filtered the solid and dried to yield (S)-l- oc-3-methylsulfonyloxy piperidine (265 g; 96.0%).
- Triethylamine (120.7 g) was added to the reaction mixture of (S)-l-Boc-3- hydroxypiperidin (200 g) and methanesulfonyl chloride (125.3g) in methylenechloride (2000 ml) at 0°C.
- the reaction mass was allowed to ambient temperature and stirred for 30 min and washed with water (2 x 2 1).
- the organic layer was concentrated, stirred with mixture of methylene chloride (120.0 ml) and hexane (1.4 1), filtered the solid and dried to yield (S)-l- oc-3-methylsulfonyloxy piperidine (254.0 g; 91.2%).
- the reaction mixture was stirred for 4-6 h at 45°C, separated the aqueous layer and concentrated under vacuum.
- the concentrated mass was stirred with methanol (225 ml) and ethyl acetate (975 ml), filtered the solid and dried.
- the resulting dried product was dissolved in DM water (800 ml) and added to sodium carbonate (55.4 g) solution and Ethyl acetate (160 ml) mixture.
- the product was filtered, washed with DM water and then the wet product was dissolved in Con.HCl (69 ml) and methanol (375 ml) mixture.
- the resulting reaction mass was concentrated, stirred with methanol (255 ml) and ethyl acetate (425 ml).
- Example-6 PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l- (PIPERIDIN-3-YL-lH-PYRAZOLO [3, 4-d] PYRIMIDINE-4- AMINE (IBRUTINIB)
- Example-8 PREPARATION OF l-[(3R)-3-[4-AMINO-3-(4- PHENOXYPHENYL)-PYRAZOLO-[3,4-d]PYRIMIDIN-l-YL]-l- PIPERIDYL]PROP-2-EN-l-ONE) (IBRUTINIB)
- reaction mass was stirred for 10 min at -85°C and washed with aqueous citric acid solution (2X500 ml; 5% w/w) followed by aqueous sodium bicarbonate solution (500 ml; 2% w/w) and water (500 ml).
- aqueous citric acid solution (2X500 ml; 5% w/w) followed by aqueous sodium bicarbonate solution (500 ml; 2% w/w) and water (500 ml).
- the organic layer was concentrated to yield syrupy mass and dissolved in MDC (50 ml) and then added Toluene (400 ml) to crystallize product.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Nitrogen Condensed Heterocyclic Rings (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
Abstract
The present invention relates to an improved process for the preparation of Ibrutinib with high purity and high yields. The present process is cost effective and feasible in large scale production also. The present process avoids the mitsunobu reagent conditions also. The present Invention also relates to a process for the preparation of Crystalline form A and Crystalline form C of Ibrutinib.
Description
AN IMPROVED PROCESS FOR THE PREPARATION OF IBRUTINIB
Field of the Invention:
The present invention relates to an improved process for the preparation of Ibrutinib with high purity and high yields. The present Invention also relates to a process for the preparation of Crystalline form A and form C of Ibrutinib.
Background of the Invention:
Ibrutinib is a kinase inhibitor indicated for the treatment of mantle cell lymphoma, chronic lymphocytic leukaemia and Waldenstrom's macroglobulinemia.
Ibrutinib is chemically is known as l-[(3R)-3-[4-amino-3-(4-phenoxyphenyl)-lH- pyrazolo[3,4-d]pyrimidin-l-yl]-l-piperidinyl]-2-propen-l-one and structurally represented as below.

IBRUTINIB
Formula-I
Ibrutinib is first disclosed in US 7514444 and marketed as IMBRUVICA ®. US '444 patent is disclosed the Ibrutinib process, wherein 4-amino-3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine is reacted with tert-Butyl 3- hydroxypiperidine-l-carboxylate and diisopropyl diazodicarboxylate to obtain intermediate under mitsunobu reaction conditions by converting the hydroxy moiety of the tert-Butyl 3-hydroxypiperidine-l-carboxylate to a better leaving group, thereby allowing a substitution reaction. Further it is converted to ibrutinib
by deprotection of Boc and acylation with acryl chloride. The reaction steps illustrated by the following scheme I:

Ibrutinib
Formula I
US 9156847 patent has disclosed another process for the preparation of ibrutinib intermediate, wherein the pyrazolo [3,4-D] pyriidine-4-amine ring of ibrutinib is preared through cyclization with use of formamide acetate and the reaction steps are illustrated in the below scheme II:
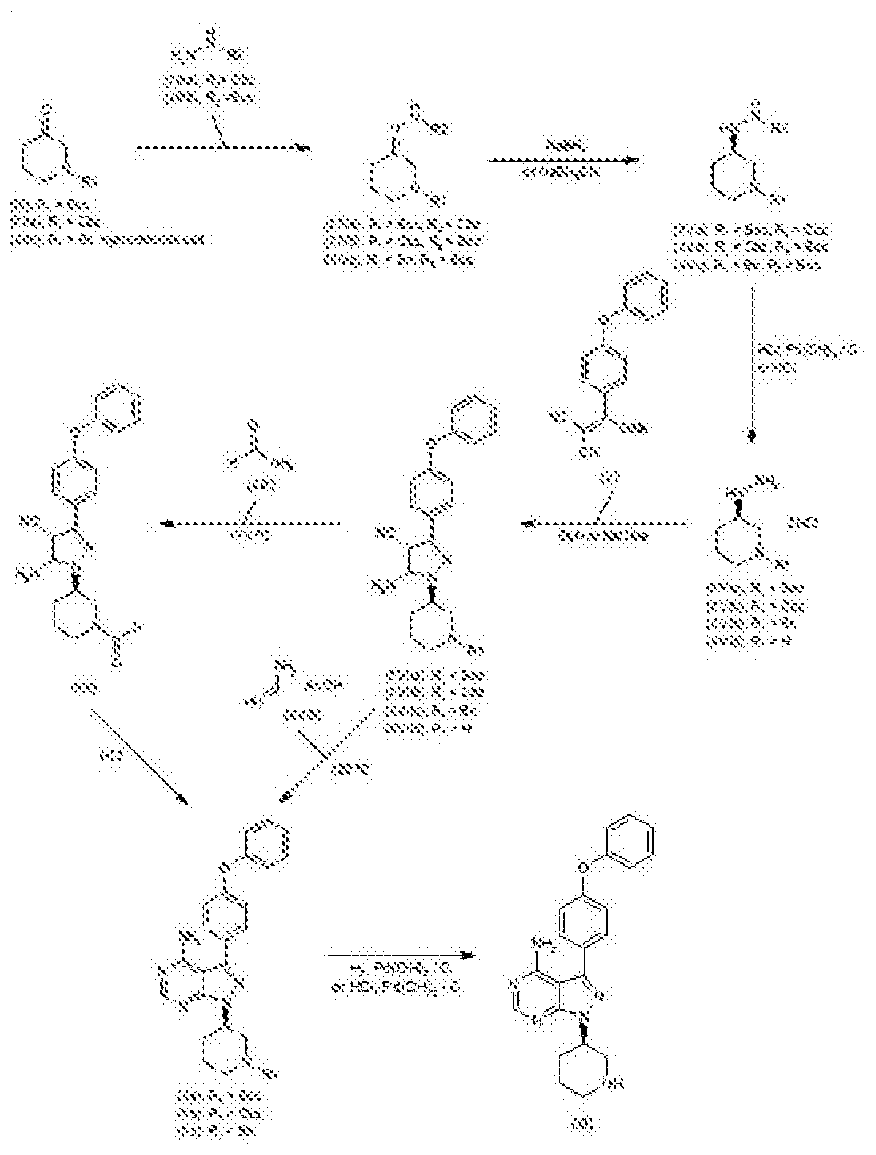
According to this patent the process which is disclosed in US 7514444 patent has a number of disadvantages, such as those associated with cost, efficiency and environmental disadvantages. For instance the Mitsunobu step may be wasteful, costly and cumbersome.
US9296753 patent has disclosed the crystalline forms A, B, C, D, E and F and their process for the preparation.
The present inventors of the present invention has developed an improved process for the preparation of Ibrutinib with high yield and high purity. The present process is cost effective and feasible in large scale production also. The present process avoids the mitsunobu reagent conditions also.
Summary of the present Invention:
One aspect of the present invention is to provide a process for the preparation of ibrutinib comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
Another aspect of the present Invention is to provide a process for the preparation of Ibrutinib Crystalline form A comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
e) dissolving ibrutinib in polar aprotic solvent and
f) isolating Crystalline form A of ibrutinib.
Yet another aspect of the present Invention is to provide a process for preparation of Ibrutinib Crystalline form C comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
e) dissolving ibrutinib in polar protic solvent and
f) isolating Crystalline form A of ibrutinib.
Yet another aspect of the present invention is to provide a purification of ibrutinib comprising the steps of:
a) dissolving ibrutinib in non-polar solvent or polar aprotic solvent or mixture thereof and
b) isolating pure ibrutinib.
Yet another aspect of the present invention is to provide a process for the preparation of Crystalline form A of Ibrutinib comprising the steps of:
dissolving ibrutinib in polar aprotic solvent and
isolating Crystalline form A of ibrutinib.
Yet another aspect of the present invention is to provide a process for preparation of Crystalline form C of Ibrutinib comprising the steps of:
dissolving ibrutinib in polar protic solvent and
isolating Crystalline form C of ibrutinib.
Yet another aspect of the present invention is to provide a process for purification of ibrutinib comprising the steps of:
a) dissolving ibrutinib in methylene chloride and toluene, and
b) isolating pure ibrutinib.
Detailed description of the Invention:
The present invention is related to an improved process for the preparation of Ibrutinib, wherein reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine and treating (S)-l-boc-3- methylsulfonyloxy piperidine with 3-(4-phenoxyphenyl)-lH-pyrazolo[3,4- d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine- 4-amine hydrochloride and converted to ibrutinib reacting with acryloyl chloride. Further ibrutinib is converted to crystalline form A and crystalline form C.
One embodiment of the present invention is to provide a process for the preparation of ibrutinib comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
The present invention is shown in the below scheme:

3-(4-phenoxyphenyl)-l-[(3R)- 3-piperidyl]pyrazolo
[3,4-d]pyrimidin-4-amine Hydro chloride
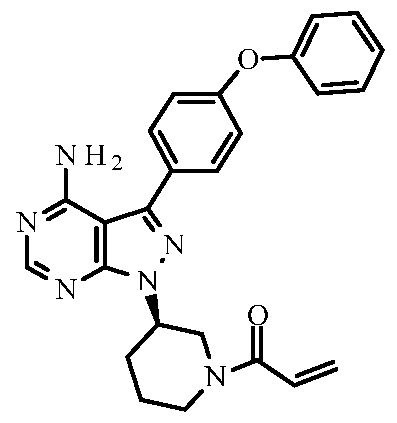
l-[(3R)-3-[4-amino-3-(4-phenoxyphenyl)pyrazolo
[3,4-d]pyriniidin-l-yl]-l-piperidyl]prop-2-en-l-one
Ibrutinib
Formula I
According to the present invention, triethylamine is added to the reaction mixture of (S)-l-Boc-3-hydroxypiperidin and methanesulfonyl chloride in methylenechloride at -10 to 10°C. The reaction mass is allowed to ambient temperature and stirred for 30 min and washed with water The organic layer is concentrated, stirred with hexane, filtered the solid and dried to yield (S)-l-boc-3- methylsulfonyloxy piperidine.
The solution of (S)-l-boc-3-methylsulfonyloxy piperidine in dimethylacetamide is added to the reaction mixture of 3-(4-phenoxyphenyl)-lH-pyrazolo [3, 4-d]
pyrimidine-4-amine, inorganic base and another inorganic base in N-methyl pyrrolidine at 45-60°C. The reaction mass is heated to 75-90°C, stirred for 14h, filtered and concentrated. The concentrated mass is dissolved in 1 : 1 mixture of methanol-toluene and added hydrochloric acid. The reaction mixture is stirred for 4-6 h at 35-50°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass is stirred with methanol and ethyl acetate, filtered the solid and dried. The resulting dried product is dissolved in DM water and added to sodium carbonate solution and Ethyl acetate mixture. The product is filtered, washed with DM water and then the wet product is dissolved in Con.HCl and methanol mixture. The resulting reaction mass is concentrated, stirred with methanol and ethyl acetate The product is filtered, washed and dried to to yield (R)-3-(4-phenoxyphenyl)-l- (piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride.
The solution of (S)-l-boc-3-methylsulfonyloxy piperidine in dimethylacetamide is added to the reaction mixture of 3-(4-phenoxyphenyl)-lH-pyrazolo [3, 4- d]pyrimidine-4-amine, inorganic base and another inorganic base in dimethylacetamide at 45-60°C. The reaction mass is heated to 75-90°C, stirred for 14h, filtered and concentrated. The concentrated mass is dissolved in 1 : 1 mixture of methanol-toluene and added hydrochloric acid. The reaction mixture is stirred for 4-6 h at 35-50°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass is stirred with methanol and ethyl acetate, filtered the solid and dried to yield (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride.
The solution of (S)-l-boc-3-methylsulfonyloxy piperidine in dimethylacetamide is added to the reaction mixture of 3-(4-phenoxyphenyl)-lH-pyrazolo [3, 4-d] pyrimidine-4-amine, inorganic base in dimethylacetamide at 45-55°C. The reaction mass is heated to 80-90°C, stirred for 14h, filtered and concentrated. The concentrated mass is dissolved in 1 : 1 mixture of methanol-toluene and added hydrochloric acid. The reaction mixture is stirred for 5-6 h at 40-50°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass is stirred
with methanol and ethyl acetate, filtered the solid and dried to yield (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride.
Acryloyl chloride is dissolved in methylene chloride and added to the cold reaction mixture of (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride and base in methylene chloride at -75 to -85°C.
The reaction mass is stirred for 10 min at -80 to -90°C and washed with aqueous citric acid solution followed by aqueous sodium bicarbonate solution and water.
The organic layer is concentrated, stirred with ethyl acetate and hexane, filtered the solid and dried to yield Ibrutinib.
Acryloyl chloride is dissolved in methylene chloride and added to the cold reaction mixture of (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride and base in methylene chloride at -75 to -85°C. The reaction mass is stirred for 10 min at -80 to -90°C and washed with aqueous citric acid solution followed by aqueous sodium bicarbonate solution and water. The organic layer is concentrated, to yield syrupy mass and dissolved in MDC and then added Toluene to crystallize product. The solid product is filtered and dried to yield Ibrutinib.
According to the present invention, Inorganic bases is selected from cesium carbonate, potassium carbonate, sodium carbonate, sodium bicarbonate, sodium phosphate, potassium phosphate, potassium phosphate, sodium acetate, potassium acetate, preferably potassium carbonate and cesium carbonate.
According to the present invention, base employed in step c) is selected from diisopropylethylamine and trimethylamine, preferably diisopropylethylamine.
Another embodiment of the present invention is to provide a purification of ibrutinib comprising the steps of:
a) dissolving ibrutinib in non-polar solvent or polar aprotic solvent or mixture thereof and
b) isolating pure ibrutinib.
According to the present invention, non-polar solvent is selected from pentane, hexane, toluene and cyclophexane, preferably hexane.
According to the present invention, polar aprotic solvent is selected from tetrahydrofuran, ethyl acetate and dimethyl formamide, preferably ethylacetate.
Yet another embodiment of the present Invention is to provide a process for the preparation of Ibrutinib Crystalline form A comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
e) dissolving ibrutinib in polar aprotic solvent and
f) isolating Crystalline form A of ibrutinib.
Yet another embodiment of the present invention is to provide a process for the preparation of Crystalline form A of Ibrutinib comprising the steps of:
a) dissolving ibrutinib in polar aprotic solvent and
b) isolating Crystalline form A of ibrutinib.
According to the present invention, polar aprotic solvent is selected from methylene chloride, ethyl acetate, acetone preferably ethyl acetate.
Yet another embodiment of the present Invention is to provide a process for the preparation of Ibrutinib Crystalline form C comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
e) dissolving ibrutinib in polar protic solvent and
f) isolating Crystalline form A of ibrutinib.
Yet another embodiment of the present invention is to provide a process for the preparation of Crystalline form C of Ibrutinib comprising the steps of:
a) dissolving ibrutinib in polar protic solvent and
b) isolating Crystalline form C of ibrutinib.
According to the present invention, polar protic solvent is selected from methanol, ethanol, n-propanol preferably methanol.
Yet another embodiment of the present invention is to provide a process for the purification of ibrutinib comprising the steps of:
a) dissolving ibrutinib in methylene chloride and toluene, and
b) isolating pure ibrutinib.
The following examples are provided for illustrative purpose only and are not intended to limit the scope of invention in anyway.
Example-1: PREPARATION OF (S)-l-boc-3-METHYLSULFONYLOXY PIPERIDINE
Triethylamine (120.7 g) was added to the reaction mixture of (S)-l-Boc-3- hydroxypiperidin (200 g) and methanesulfonyl chloride (125.3g) in methylenechloride (2000 ml) at 0°C. The reaction mass was allowed to ambient temperature and stirred for 30 min and washed with water (2 x 2 1). The organic layer was concentrated, stirred with hexane (1.4 1), filtered the solid and dried to yield (S)-l- oc-3-methylsulfonyloxy piperidine (265 g; 96.0%).
Example-2: PREPARATION OF (S)-l-boc-3-METHYLSULFONYLOXY PIPERIDINE
Triethylamine (120.7 g) was added to the reaction mixture of (S)-l-Boc-3- hydroxypiperidin (200 g) and methanesulfonyl chloride (125.3g) in methylenechloride (2000 ml) at 0°C. The reaction mass was allowed to ambient temperature and stirred for 30 min and washed with water (2 x 2 1). The organic layer was concentrated, stirred with mixture of methylene chloride (120.0 ml) and hexane (1.4 1), filtered the solid and dried to yield (S)-l- oc-3-methylsulfonyloxy piperidine (254.0 g; 91.2%).
HPLC Purity: 99.5%; chiral purity: 100%
Example-3: PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l- (PIPERIDIN-3-YL-lH-PYRAZOLO[3,4-d]PYRIMIDINE-4- AMINE
HYDROCHLORIDE
The solution of (S)-l- oc-3-methylsulfonyloxy piperidine (173.3 g) in N-methyl- 2-pyrrolidone (525 ml) was added to the reaction mixture of 3-(4-phenoxyphenyl)- lH-pyrazolo[3,4-d]pyrimidine-4-amine (75 g), cesium carbonate (217.7 g) in N-
methyl-2-pyrrolidone (600 ml) at 55°C. The reaction mass was heated to 80-85°C, stirred for 6-8h, water workup and partially concentrated. The partial concentrated mass was dissolved in methanol (935 ml) and added hydrochloric acid (186 ml; 18% w/v). The reaction mixture was stirred for 4-6 h at 45°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass was stirred with methanol (225 ml) and ethyl acetate (975 ml), filtered the solid and dried. The resulting dried product was dissolved in DM water (800 ml) and added to sodium carbonate (55.4 g) solution and Ethyl acetate (160 ml) mixture. The product was filtered, washed with DM water and then the wet product was dissolved in Con.HCl (69 ml) and methanol (375 ml) mixture. The resulting reaction mass was concentrated, stirred with methanol (255 ml) and ethyl acetate (425 ml). The product was filtered, washed and dried to yield (R)-3-(4-phenoxyphenyl)-l- (piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride (69 g; 60.7%).
HPLC Purity: 99.9%; chiral purity: 99.9%
Example-4: PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l-
(PIPERIDIN-3-YL-lH-PYRAZOLO[3,4-d]PYRIMIDINE-4-AMINE
HYDROCHLORIDE
The solution of (S)-l- oc-3-methylsulfonyloxy piperidine (115.5 g) in dimethylacetamide (300 ml) was added to the reaction mixture of 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine (50 g), cesium carbonate (13.4 g) and potassium carbonate (108.3 g) in dimethylacetamide (450 ml) at 55°C. The reaction mass was heated to 85°C, stirred for 14h, filtered and concentrated. The concentrated mass was dissolved in 1 : 1 mixture of methanol-toluene (1250 ml) and added hydrochloric acid (117 ml; 18% w/v). The reaction mixture was stirred for 6 h at 45°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass was stirred with methanol (100 ml) and ethyl acetate (625 ml), filtered the solid and dried to yield (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH- pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride (59 g; 78.4%).
HPLC Purity: 99.6%
Example-5: PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l-
(PIPERIDIN-3-YL-lH-PYRAZOLO[3,4-d]PYRIMIDINE-4-AMINE
HYDROCHLORIDE
The solution of (S)-l- oc-3-methylsulfonyloxy piperidine (11.55 g) in dimethylacetamide (20 ml) was added to the reaction mixture of 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine (5 g) and potassium carbonate (6.15 g) in dimethylacetamide (55 ml) at 55°C. The reaction mass was heated to 100°C, stirred for 28h, filtered and concentrated. The concentrated mass was dissolved in 1 : 1 mixture of methanol -toluene (86 ml) and added hydrochloric acid (86 ml; 18% w/v). The reaction mixture was stirred for 30 h at 25°C, separated the aqueous layer and concentrated under vacuum. The concentrated mass was stirred with ethyl acetate (65 ml), filtered the solid and dried to yield (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride (3.5 g; 46.1%).
HPLC Purity: 98.5%
Example-6: PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l- (PIPERIDIN-3-YL-lH-PYRAZOLO [3, 4-d] PYRIMIDINE-4- AMINE (IBRUTINIB)
Acryloyl chloride (9.85 g) was dissolved in methylene chloride (150 ml) and added to the cold reaction mixture of (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH- pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride (50 g) and diisopropylethylamine (63.2 g) in methylene chloride (1000 ml) at -85°C. The reaction mass was stirred for 10 min at -85°C and washed with aqueous citric acid solution (500 ml; 5% w/w) followed by aqueous sodium bicarbonate solution (500 ml; 5% w/w) and water (500 ml). The organic layer was concentrated, stirred with ethyl acetate (300 ml) and hexane (300 ml), filtered the solid and dried to yield (R)- 3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine crude (40.6 g; 84.8%).
HPLC Purity: 99.7%
Example-7: PREPARATION OF (R)-3-(4-PHENOXYPHENYL)-l- (PIPERIDIN-3-YL-lH-PYRAZOLO [3, 4-d] PYRIMIDINE-4- AMINE (IBRUTINIB)
Acryloyl chloride (1 g) was dissolved in methylene chloride (10 ml) and added to the cold reaction mixture of (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH- pyrazolo [3, 4-d] pyrimidine-4-amine hydrochloride (5 g) and diisopropylethylamine (5.6 g) in methylene chloride (75 ml) at -45°C. The reaction mass was stirred for 30 min at -45°C and washed with aqueous citric acid solution (50 ml; 5% w/w) followed by aqueous sodium bicarbonate solution (50 ml; 5% w/w) and water (50 ml). The organic layer was concentrated, stirred with ethyl acetate (30 ml), filtered the solid and dried to yield (R)-3-(4-phenoxyphenyl)-l- (piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine crude (2.9 g; 60.6%).
Example-8:PREPARATION OF l-[(3R)-3-[4-AMINO-3-(4- PHENOXYPHENYL)-PYRAZOLO-[3,4-d]PYRIMIDIN-l-YL]-l- PIPERIDYL]PROP-2-EN-l-ONE) (IBRUTINIB)
Acryloyl chloride (10.34 g) was dissolved in methylene chloride (50 ml) and added to the cold reaction mixture of (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH- pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride (50 g) and diisopropylethylamine (63.3 g) in methylene chloride (1000 ml) at -85°C. The reaction mass was stirred for 10 min at -85°C and washed with aqueous citric acid solution (2X500 ml; 5% w/w) followed by aqueous sodium bicarbonate solution (500 ml; 2% w/w) and water (500 ml). The organic layer was concentrated to yield syrupy mass and dissolved in MDC (50 ml) and then added Toluene (400 ml) to crystallize product. The solid product was filtered and dried to yield l-[(3R)-3-[4- Amino-3-(4-Phenoxyphenyl)-Pyrazolo-[3,4-d]Pyrimidin-l-Yl]-l-Piperidyl]Prop- 2-en-l-one (51 g)
HPLC Purity: 99.8%; chiral purity: 99.9%.
Example-9: PREPARATION IBRUTINIB FORM - A
1 - [(3R)-3 - [4- Amino-3 -(4-Phenoxyphenyl)-Pyrazolo-[3 ,4-d]Pyrimidin- 1 - Yl]- 1 - Piperidyl]Prop-2-en-l-one (5.0 g), was dissolved in ethyl acetate (35 ml) at 25- 35°C, filtered and crystallized from ethyl acetate. The product was filtered, washed and dried to yield pure Ibrutinib crystalline Form-A (3.3 g)
Example-10: PREPARATION IBRUTINIB PHARMA FORM - C
1 - [(3R)-3 - [4- Amino-3 -(4-Phenoxyphenyl)-Pyrazolo-[3 ,4-d]Pyrimidin- 1 - Yl]- 1 - Piperidyl]Prop-2-en-l-one (70.0 g) was dissolved in methanol (875 ml) at 50-55°C, filtered and crystallized from methanol. The product was filtered, washed and dried to yield pure Ibrutinib crystalline Form-C (51 g; 72.8%).
HPLC Purity: 99.68%; chiral purity: 99.9%.
Claims
1. A process for the preparation of Ibrutinib comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib.
A process for the preparation of Ibrutinib Crystalline form A comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piperidine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib,
e) dissolving ibrutinib in polar aprotic solvent and
f) isolating Crystalline form A of ibrutinib.
A process for the preparation of Ibrutinib Crystalline form C comprising the steps of:
a) reacting (S)-l-boc -3 -hydroxy piped dine with methane sulfonyl chloride to get (S)-l-boc-3-methylsulfonyloxy piperidine,
b) treating (S)-l-boc-3-methylsulfonyloxy piperidine with 3-(4- phenoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidine-4-amine in presence of inorganic base or mixture of inorganic bases to get (R)-3-(4- phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4-d]pyrimidine-4-amine hydrochloride,
c) reacting (R)-3-(4-phenoxyphenyl)-l-(piperidin-3-yl-lH-pyrazolo[3,4- d]pyrimidine-4-amine hydrochloride with base followed by acryloyl chloride to obtain ibrutinib,
d) optionally purifying the ibrutinib,
e) dissolving ibrutinib in polar protic solvent and
f) isolating Crystalline form C of ibrutinib.
A process for the preparation of Crystalline form A of Ibrutinib comprising the steps of:
a) dissolving ibrutinib in polar aprotic solvent and
b) isolating Crystalline form A of ibrutinib.
A process for the preparation of Crystalline form C of Ibrutinib comprising the steps of:
a) dissolving ibrutinib in polar protic solvent and
b) isolating Crystalline form C of ibrutinib.
A process according to claims 1,2 and 3, wherein, Inorganic base is selected from cesium carbonate, potassium carbonate, sodium carbonate, sodium bicarbonate, sodium phosphate, potassium phosphate, potassium phosphate, sodium acetate, potassium acetate.
7. A process according to claims 1, 2 and 3, wherein, base employed in step c) is selected from diisopropylethylamine and trimethylamine.
8. A process according to claims 2 and 4, wherein, polar aprotic solvent is selected from methylene chloride, ethyl acetate.
9. A process according claims 3 and 5, wherein, polar protic solvent is selected from methanol, ethanol, n-propanol.
10. A process for the purification of ibrutinib comprising the steps of:
a) dissolving ibrutinib in methylene chloride and toluene, and
b) isolating pure ibrutinib.
Priority Applications (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US16/074,505 US10626117B2 (en) | 2016-02-01 | 2017-02-01 | Process for the preparation of Ibrutinib |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| IN201641003475 | 2016-02-01 | ||
| IN201641003475 | 2016-02-01 |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| WO2017134684A2 true WO2017134684A2 (en) | 2017-08-10 |
| WO2017134684A3 WO2017134684A3 (en) | 2017-10-05 |
Family
ID=58537050
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/IN2017/050045 Ceased WO2017134684A2 (en) | 2016-02-01 | 2017-02-01 | An improved process for the preparation of ibrutinib |
Country Status (2)
| Country | Link |
|---|---|
| US (1) | US10626117B2 (en) |
| WO (1) | WO2017134684A2 (en) |
Cited By (5)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2019195827A1 (en) * | 2018-04-06 | 2019-10-10 | Johnson Matthey Public Limited Company | Novel form of ibrutinib |
| WO2019213184A1 (en) | 2018-05-03 | 2019-11-07 | Juno Therapeutics, Inc. | Combination therapy of a chimeric antigen receptor (car) t cell therapy and a kinase inhibitor |
| CN112251478A (en) * | 2020-10-26 | 2021-01-22 | 宁波酶赛生物工程有限公司 | Ketoreductase and enzyme catalysis preparation method of S-1-BOC-3 hydroxypiperidine |
| WO2022157250A1 (en) | 2021-01-21 | 2022-07-28 | Synthon B.V. | Process for making ibrutinib |
| WO2023220655A1 (en) | 2022-05-11 | 2023-11-16 | Celgene Corporation | Methods to overcome drug resistance by re-sensitizing cancer cells to treatment with a prior therapy via treatment with a t cell therapy |
Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US7514444B2 (en) | 2006-09-22 | 2009-04-07 | Pharmacyclics, Inc. | Inhibitors of bruton's tyrosine kinase |
| US9156847B2 (en) | 2013-03-15 | 2015-10-13 | Janssen Pharmaceutica Nv | Processes and intermediates for preparing a medicament |
| US9296753B2 (en) | 2012-06-04 | 2016-03-29 | Pharmacyclics Llc | Crystalline forms of a Bruton's tyrosine kinase inhibitor |
Family Cites Families (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| DK2710007T3 (en) | 2011-05-17 | 2020-01-27 | Principia Biopharma Inc | kinase inhibitors |
| US8841449B2 (en) | 2011-11-09 | 2014-09-23 | Vertex Pharmaceuticals Incorporated | Compounds useful as inhibitors of ATR kinase |
-
2017
- 2017-02-01 WO PCT/IN2017/050045 patent/WO2017134684A2/en not_active Ceased
- 2017-02-01 US US16/074,505 patent/US10626117B2/en active Active
Patent Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US7514444B2 (en) | 2006-09-22 | 2009-04-07 | Pharmacyclics, Inc. | Inhibitors of bruton's tyrosine kinase |
| US9296753B2 (en) | 2012-06-04 | 2016-03-29 | Pharmacyclics Llc | Crystalline forms of a Bruton's tyrosine kinase inhibitor |
| US9156847B2 (en) | 2013-03-15 | 2015-10-13 | Janssen Pharmaceutica Nv | Processes and intermediates for preparing a medicament |
Cited By (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2019195827A1 (en) * | 2018-04-06 | 2019-10-10 | Johnson Matthey Public Limited Company | Novel form of ibrutinib |
| WO2019213184A1 (en) | 2018-05-03 | 2019-11-07 | Juno Therapeutics, Inc. | Combination therapy of a chimeric antigen receptor (car) t cell therapy and a kinase inhibitor |
| CN112251478A (en) * | 2020-10-26 | 2021-01-22 | 宁波酶赛生物工程有限公司 | Ketoreductase and enzyme catalysis preparation method of S-1-BOC-3 hydroxypiperidine |
| CN112251478B (en) * | 2020-10-26 | 2022-08-30 | 宁波酶赛生物工程有限公司 | Ketoreductase and enzyme catalysis preparation method of S-1-BOC-3 hydroxypiperidine |
| WO2022157250A1 (en) | 2021-01-21 | 2022-07-28 | Synthon B.V. | Process for making ibrutinib |
| WO2023220655A1 (en) | 2022-05-11 | 2023-11-16 | Celgene Corporation | Methods to overcome drug resistance by re-sensitizing cancer cells to treatment with a prior therapy via treatment with a t cell therapy |
Also Published As
| Publication number | Publication date |
|---|---|
| WO2017134684A3 (en) | 2017-10-05 |
| US10626117B2 (en) | 2020-04-21 |
| US20190241567A1 (en) | 2019-08-08 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| JP6921087B2 (en) | Ruxolitinib synthesis process | |
| US9527855B2 (en) | Process for preparing chiral dipeptidyl peptidase-IV inhibitors | |
| JP6523566B2 (en) | Crystal form of dihydropyrido ring compound, production method and intermediate | |
| US10626117B2 (en) | Process for the preparation of Ibrutinib | |
| US8981105B2 (en) | Process of preparing a thrombin specific inhibitor | |
| JP5735659B2 (en) | Method for producing sitagliptin intermediate | |
| US20080154045A1 (en) | Process for Producing 1-Substituted-3-Fluoroalkylpyrazole-4-Carboxylate | |
| EP2609099A2 (en) | Sitagliptin, salts and polymorphs thereof | |
| WO2016035007A2 (en) | An improved process for the preparation of apixaban and intermediates thereof | |
| WO2017163257A1 (en) | Process for preparing pure lh-pyrazolo[3,4-d] pyrimidine derivative | |
| US9540344B2 (en) | Production method for isoquinoline derivatives and salts thereof | |
| CN107814804A (en) | The preparation method of Buddhist nun is replaced according to Shandong | |
| IT201800006337A1 (en) | LIFITEGRAST PREPARATION PROCEDURE | |
| CN119032095A (en) | Improved method for preparing ruxolitinib | |
| CN108623602A (en) | A method of prepare and purify and replaces Buddhist nun according to Shandong | |
| US20190040066A1 (en) | Process for the preparation of phosphatidylinositol 3-kinase inhibitor | |
| US20120010405A1 (en) | Optically active 3-amino-2,5-dioxopyrrolidine-3-carboxylate, process for production of the compound, and use of the compound | |
| US8093384B2 (en) | Processes for the preparation of alfuzosin | |
| CA2431821C (en) | Method for the production of 2-(2-ethoxyphenyl)-substituted imidazotriazinones | |
| JP7577743B2 (en) | Processes and intermediates for the manufacture of tofacitinib | |
| RU2810976C1 (en) | Method for producing tofacitinib and its intermediates | |
| CN103649078B (en) | The phenyl compound replaced | |
| CN119161348A (en) | Ibrutinib and its synthesis method | |
| CN119899173A (en) | A method for preparing nitroso-ceritinib containing genotoxic impurity of ceritinib | |
| CN102443032B (en) | Method for preparing antiviral composition and application of its intermediate |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 17717001 Country of ref document: EP Kind code of ref document: A2 |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| 122 | Ep: pct application non-entry in european phase |
Ref document number: 17717001 Country of ref document: EP Kind code of ref document: A2 |