WO2011048425A1 - Continuous process for the production of beta-keto esters by claisen condensation - Google Patents
Continuous process for the production of beta-keto esters by claisen condensation Download PDFInfo
- Publication number
- WO2011048425A1 WO2011048425A1 PCT/GB2010/051778 GB2010051778W WO2011048425A1 WO 2011048425 A1 WO2011048425 A1 WO 2011048425A1 GB 2010051778 W GB2010051778 W GB 2010051778W WO 2011048425 A1 WO2011048425 A1 WO 2011048425A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- compound
- reaction zone
- process according
- formula
- compounds
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07B—GENERAL METHODS OF ORGANIC CHEMISTRY; APPARATUS THEREFOR
- C07B41/00—Formation or introduction of functional groups containing oxygen
- C07B41/06—Formation or introduction of functional groups containing oxygen of carbonyl groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C253/00—Preparation of carboxylic acid nitriles
- C07C253/30—Preparation of carboxylic acid nitriles by reactions not involving the formation of cyano groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C255/00—Carboxylic acid nitriles
- C07C255/01—Carboxylic acid nitriles having cyano groups bound to acyclic carbon atoms
- C07C255/19—Carboxylic acid nitriles having cyano groups bound to acyclic carbon atoms containing cyano groups and carboxyl groups, other than cyano groups, bound to the same saturated acyclic carbon skeleton
- C07C255/21—Carboxylic acid nitriles having cyano groups bound to acyclic carbon atoms containing cyano groups and carboxyl groups, other than cyano groups, bound to the same saturated acyclic carbon skeleton the carbon skeleton being further substituted by doubly-bound oxygen atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C67/00—Preparation of carboxylic acid esters
- C07C67/30—Preparation of carboxylic acid esters by modifying the acid moiety of the ester, such modification not being an introduction of an ester group
- C07C67/313—Preparation of carboxylic acid esters by modifying the acid moiety of the ester, such modification not being an introduction of an ester group by introduction of doubly bound oxygen containing functional groups, e.g. carboxyl groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C67/00—Preparation of carboxylic acid esters
- C07C67/30—Preparation of carboxylic acid esters by modifying the acid moiety of the ester, such modification not being an introduction of an ester group
- C07C67/333—Preparation of carboxylic acid esters by modifying the acid moiety of the ester, such modification not being an introduction of an ester group by isomerisation; by change of size of the carbon skeleton
- C07C67/343—Preparation of carboxylic acid esters by modifying the acid moiety of the ester, such modification not being an introduction of an ester group by isomerisation; by change of size of the carbon skeleton by increase in the number of carbon atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C69/00—Esters of carboxylic acids; Esters of carbonic or haloformic acids
- C07C69/66—Esters of carboxylic acids having esterified carboxylic groups bound to acyclic carbon atoms and having any of the groups OH, O—metal, —CHO, keto, ether, acyloxy, groups, groups, or in the acid moiety
- C07C69/67—Esters of carboxylic acids having esterified carboxylic groups bound to acyclic carbon atoms and having any of the groups OH, O—metal, —CHO, keto, ether, acyloxy, groups, groups, or in the acid moiety of saturated acids
- C07C69/716—Esters of keto-carboxylic acids or aldehydo-carboxylic acids
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D319/00—Heterocyclic compounds containing six-membered rings having two oxygen atoms as the only ring hetero atoms
- C07D319/04—1,3-Dioxanes; Hydrogenated 1,3-dioxanes
- C07D319/06—1,3-Dioxanes; Hydrogenated 1,3-dioxanes not condensed with other rings
Definitions
- the present invention concerns a process for the production of certain pharmaceutically useful intermediate compounds, in particular (5R)-1 ,1- dimethylethyl-6-cyano-5-hydroxy-3-oxo-hexanoate.
- (5R)-1 1-dimethylethyl-6-cyano-5-hydroxy-3-oxo-hexanoate is a useful pharmaceutical intermediate particularly in the manufacture of statin drugs such as atorvastatin, sold under the trade name LipitorTM.
- Tertiary butyl acetate enolate decomposes to the ketene which then reacts with another molecule of tert-butyl acetate enolate to self condense to give tert-butylacetoacetate.
- tert-Butylacetoacetate is the major impurity in all Claisen type reactions involving TBA. Since the reagents that go to make tert-butyl acetate enolate, in particular the lithium amide base, are expensive, the formation of tert-butylacetoacetate is a costly inefficiency.
- R is a straight or branched chain alkyl group
- R 1 is a straight or branched chain alkyl group substituted with a nitrile group, a hydroxy group or a halogen atom;
- R 2 is a hydroxy group or a keto group
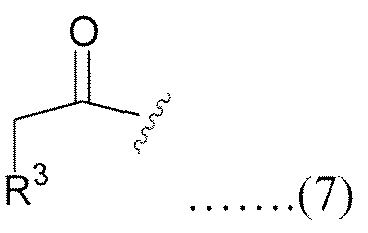
- each R 3 is, independently, hydrogen or a straight or branched chain alkyl group, the process comprising providing to a reaction zone a continuous stream of a compound of formula (3): wherein R and R 3 are as previously defined and R 4 is hydrogen or has the general formula (7):
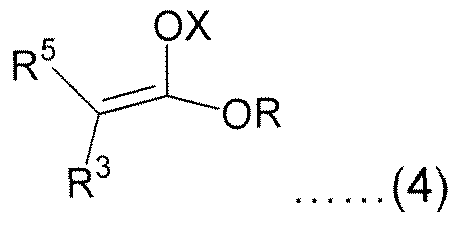
- R 3 is as defined above and a continuous stream of an alkali metal or alkaline earth metal amide base, alkyl lithium or Grignard reagent; contacting the continuous streams together in the reaction zone to yield the enolate of formula (4):
- R and R 3 are as previously defined, X is an alkali metal or alkaline earth metal, and R 5 is hydrogen or has the general formula (8):
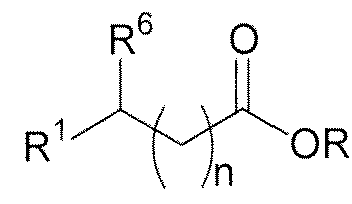
- R and R 1 are as previously defined or together define a ring structure
- R 6 is hydrogen, hydroxyl, alkoxyl or a keto group and n is 0 or 1 ; and contacting the continuous stream of compound (5) with a continuous stream of the enolate (4) in the or the separate reaction zone at a temperature above 20°C to yield a compound of formula (1 ): wherein R 1 , R and X are as previously defined, and treating the compound of formula (1 ) with an acid to yield the compound of formula (6).
- enolates of the type represented by formula (4) are prepared at low temperature due to their thermal instability.
- the reaction between compound (4) and compound (5) also conventionally takes place at low temperature. This is because on an industrial scale if one prepares an 8000L batch of enolate mixture at -60 °C and one wants to carry out a subsequent reaction at 10 °C using this enolate solution it is not possible to warm this solution to 10 °C at a rate faster than the enolate will decompose. Lying behind the present invention is the realisation that it is possible prepare the enolate at a higher than conventional temperature and use it immediately also at a higher than conventional temperature.
- reaction partner for example 4-cyano-3-hydroxybutyric acid ethyl ester
- a significant advantage of the inventive process is therefore that it allows the use of non- cryogenic conditions in both the synthesis and use of ester enolates, and also for rapid production of compound (6) on an industrial scale.
- the enolisation reaction is conducted at least partially before contacting the enol compound (4) with its reaction partner compound (5).
- the steps of providing to the reaction zone a continuous stream of a compound of formula (3) and a continuous stream of an alkali metal or alkaline earth metal amide base, alkyl lithium or a Grignard reagent, and the steps of providing to the or the separate reaction zone a continuous stream of a compound of formula (5) are sequential steps in the process of the invention.
- This aspect of the invention is found to be particularly advantageous when compound (5) is itself unstable in the presence of the alkali metal or alkaline earth metal amide base, alkyl lithium or a Grignard reagent, as appears to be the case for example when R 1 contains a nitrile group.
- R 1 contains a nitrile group.
- the stoichiometric ratio of alkali metal or alkaline earth metal amide base, alkyl lithium or a Grignard reagent to compound (5) supplied to the and/or to the separate reaction zone is less than about 4.5 : 1 , more preferably less than about 4.0 : 1 and most preferably less than about 3.5 : 1 .
- Continuous flow production of the unstable compound (4) allows the compound to be used as it is formed, and allows the use of very high heat/mass transfer flow equipment, permitting excellent temperature control of the reaction mixture.
- the temperature at which compounds (4) and (5) are reacted together is above 25°C, more preferably above 30°C.
- a significant advantage of using a relatively high temperature in the reaction between compounds (4) and (5) is that only a low residence time in the or the separate reaction zone need be employed.
- the residence time of the contacted continuous streams of compounds (4) and (5) in the or the separate reaction zone is less than about 5 minutes, more preferably less than about I minute, still more preferably less than about 50 seconds and most preferably less than about 40, or even 30 seconds.
- the residence time of the contacted continuous streams of compound (3) and the alkali metal or alkaline earth metal amide base, alkyl lithium or Grignard reagent in the reaction zone is less than about 5 minutes, more preferably less than about 4 minutes, still more preferably less than about 3 minutes and most preferably less than about 2 minutes.
- the enolate compound (4) is prepared from the reaction of compound (3) and the alkali metal or alkaline earth metal amide base, alkyl lithium or Grignard reagent in a first reaction zone, and the compound of formula (1 ) is prepared from the reaction between compounds (4) and (5) in a second reaction zone.
- first and second reaction zones are preferable rather than essential in the process of the invention and, particularly when R 1 contains a halogen atom, does not appear to compromise purity of the product unduly.
- the treatment of compound (1 ) with acid may take place in the same or a different reaction zone as that in which the reaction between compounds (4) and (5) takes place, and this step of the process need not be continuous, although it can be.
- continuous is preferably meant that steady state reaction conditions prevail in the or the separate reaction zone as far as the reactions between compound (3) and the alkali metal or alkaline earth metal amide base, alkyl lithium or Grignard reagent and/or between compounds (4) and (5) are concerned.
- reagent streams may be supplied to the or the separate reaction zone, and product stream(s) may be recovered therefrom as consistent continuous streams or as intermittent or pulsed streams.
- R 1 is preferably a substituted methyl group.
- the halogen atom is preferably chlorine.
- R is preferably tertiary butyl.
- X is preferably lithium and the alkali metal or alkaline earth metal amide base is preferably a lithium amide base, such as lithium hexamethyldisilazane or lithium diiospropylamide, lithium dicyclohexylamide or lithium amide.
- a lithium amide base such as lithium hexamethyldisilazane or lithium diiospropylamide, lithium dicyclohexylamide or lithium amide.
- R 0R (7) comprising obtaining compound (6) by the aforementioned process and subjecting that compound to reducing conditions to obtain compound (7).
- the reducing conditions are at least partially provided by one or more enzymes.
- the invention also provides a process for the preparation of compound (8) as aforesaid and further converting compound (8) into a useful pharmaceutical compound.
- the invention will now be more particularly described with reference to the following example.
- t-Butyl Acetate enolate was prepared by pumping two solutions through a 1 .016mm i.d. stainless steel capillary tube:
- reaction temperature was controlled by submerging the entire capillary reactor in a Huber heater/ chiller unit with a set-point of 0°C.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Abstract
Description




Claims




Priority Applications (6)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| EP10773387A EP2491010A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
| US13/522,455 US20120309989A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
| AU2010309541A AU2010309541A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by Claisen condensation |
| CN2010800542522A CN102639489A (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
| CA2780027A CA2780027A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
| US14/063,264 US20140051869A1 (en) | 2009-10-23 | 2013-10-25 | Continuous process for the production of beta-keto esters by claisen condensation |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| GB0918613.1 | 2009-10-23 | ||
| GB0918613A GB2474687A (en) | 2009-10-23 | 2009-10-23 | A continuous process for the production of (R)-6-cyano-5-hydroxy-3-oxo-hexanoic acid tert-butyl ester (and derivatives) |
Related Child Applications (2)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| US13/522,455 A-371-Of-International US20120309989A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
| US14/063,264 Continuation US20140051869A1 (en) | 2009-10-23 | 2013-10-25 | Continuous process for the production of beta-keto esters by claisen condensation |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2011048425A1 true WO2011048425A1 (en) | 2011-04-28 |
Family
ID=41426606
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/GB2010/051778 Ceased WO2011048425A1 (en) | 2009-10-23 | 2010-10-22 | Continuous process for the production of beta-keto esters by claisen condensation |
Country Status (7)
| Country | Link |
|---|---|
| US (2) | US20120309989A1 (en) |
| EP (1) | EP2491010A1 (en) |
| CN (1) | CN102639489A (en) |
| AU (1) | AU2010309541A1 (en) |
| CA (1) | CA2780027A1 (en) |
| GB (1) | GB2474687A (en) |
| WO (1) | WO2011048425A1 (en) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2014013261A1 (en) * | 2012-07-18 | 2014-01-23 | Bakhu Pharma Limited | Process for crossed claisen condensation reactions promoted by lithium amide in liquid ammonia |
Families Citing this family (5)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN105461593B (en) * | 2015-12-31 | 2018-02-27 | 江西科苑生物药业有限公司 | A kind of continuous preparation method of the oxo hecanoic acid t-butyl ester of 6 cyano group, 5 hydroxyl 3 |
| EP3360857A1 (en) * | 2017-02-13 | 2018-08-15 | Patheon Austria GmbH Co. & KG | Process for preparing pentenoate |
| US11999757B2 (en) | 2017-11-01 | 2024-06-04 | Melinta Subsidiary Corp. | Synthesis of boronate ester derivatives and uses thereof |
| CN108033899B (en) * | 2017-12-06 | 2020-04-10 | 浙江科技学院 | Preparation method of (R) -6-cyano-5-hydroxy-3-carbonyl hexanoate tert-butyl ester |
| CN116120173A (en) * | 2023-02-22 | 2023-05-16 | 金浦新材料股份有限公司 | Synthesis method of long-chain alkyl-beta-keto ester compound |
Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0643689A1 (en) | 1991-10-11 | 1995-03-22 | Warner-Lambert Company | Improved process for the synthesis of (5r)-1,1-dimethylethyl 6-cyano-5-hydroxy-3-oxo-hexanoate |
| US6340767B1 (en) | 1999-06-04 | 2002-01-22 | Kaneka Corporation | Processes for the preparation of 5-hydroxy-3-oxopentanoic acid derivatives |
| US20030040634A1 (en) * | 1998-08-05 | 2003-02-27 | Noriyuki Kizaki | Process for producing optically active 2-[6-(hydroxymethyl)-1,3-dioxan-4yl]acetic acid derivatives |
Family Cites Families (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| SG44830A1 (en) * | 1992-07-02 | 1997-12-19 | Hoechst Ag | Process for preparing tert-butyl 93R,5S) -6 Hydroxy-3,5-O-Isopropylidene-3, 5-Dihydroxyhexanoate |
-
2009
- 2009-10-23 GB GB0918613A patent/GB2474687A/en not_active Withdrawn
-
2010
- 2010-10-22 AU AU2010309541A patent/AU2010309541A1/en not_active Abandoned
- 2010-10-22 CN CN2010800542522A patent/CN102639489A/en active Pending
- 2010-10-22 CA CA2780027A patent/CA2780027A1/en not_active Abandoned
- 2010-10-22 WO PCT/GB2010/051778 patent/WO2011048425A1/en not_active Ceased
- 2010-10-22 EP EP10773387A patent/EP2491010A1/en not_active Withdrawn
- 2010-10-22 US US13/522,455 patent/US20120309989A1/en not_active Abandoned
-
2013
- 2013-10-25 US US14/063,264 patent/US20140051869A1/en not_active Abandoned
Patent Citations (4)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0643689A1 (en) | 1991-10-11 | 1995-03-22 | Warner-Lambert Company | Improved process for the synthesis of (5r)-1,1-dimethylethyl 6-cyano-5-hydroxy-3-oxo-hexanoate |
| US20030040634A1 (en) * | 1998-08-05 | 2003-02-27 | Noriyuki Kizaki | Process for producing optically active 2-[6-(hydroxymethyl)-1,3-dioxan-4yl]acetic acid derivatives |
| US6903225B2 (en) | 1998-08-05 | 2005-06-07 | Kaneka Corporation | Process for producing optically active 2-[6-(hydroxymethyl)-1,3-dioxan-4-yl]acetic acid derivatives |
| US6340767B1 (en) | 1999-06-04 | 2002-01-22 | Kaneka Corporation | Processes for the preparation of 5-hydroxy-3-oxopentanoic acid derivatives |
Non-Patent Citations (1)
| Title |
|---|
| WILES C ET AL: "The preparation and reaction of enolates within micro reactors", TETRAHEDRON, ELSEVIER SCIENCE PUBLISHERS, AMSTERDAM, NL, vol. 61, no. 45, 7 November 2005 (2005-11-07), pages 10757 - 10773, XP025383702, ISSN: 0040-4020, [retrieved on 20051107], DOI: DOI:10.1016/J.TET.2005.08.076 * |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2014013261A1 (en) * | 2012-07-18 | 2014-01-23 | Bakhu Pharma Limited | Process for crossed claisen condensation reactions promoted by lithium amide in liquid ammonia |
Also Published As
| Publication number | Publication date |
|---|---|
| EP2491010A1 (en) | 2012-08-29 |
| AU2010309541A1 (en) | 2012-05-31 |
| CA2780027A1 (en) | 2011-04-28 |
| US20140051869A1 (en) | 2014-02-20 |
| GB0918613D0 (en) | 2009-12-09 |
| US20120309989A1 (en) | 2012-12-06 |
| CN102639489A (en) | 2012-08-15 |
| GB2474687A (en) | 2011-04-27 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| US20140051869A1 (en) | Continuous process for the production of beta-keto esters by claisen condensation | |
| KR20090066910A (en) | Improved method for preparing L-3-O-substituted-ascorbic acid | |
| KR20020068517A (en) | Crystals of penicillin and process for the production thereof | |
| US7772436B2 (en) | Process for producing 2,2′-bis(trifluoromethyl)-4,4′-diaminobiphenyl | |
| CA2278713A1 (en) | Process for producing butyric ester derivatives | |
| JPH0578277A (en) | Production of 3,3,3-trifluorolactic acid and method for improving optical purity | |
| JPS61229852A (en) | Production of 1-methyl-5-hydroxypyrazole | |
| US6573397B2 (en) | Process for producing 3,4-dihydroxybenzonitrile | |
| JP2002128763A (en) | Method for purifying 1,3-dialkylpyrazole-4-carboxylic acid | |
| US7038091B2 (en) | Process for producing acetylene compound | |
| EP1352898B1 (en) | Process for producing beta-ketonitrile compound | |
| CN114853692B (en) | Preparation method of 2-aminothiazole | |
| CN115872995B (en) | Pyrazolopyridine compound and preparation method of carboxylic acid derivative | |
| CN117105801B (en) | A method for preparing oseltamivir and its phosphate, and its intermediates. | |
| CN121202767A (en) | A method for preparing 1-(4-fluoro-3-pyridyl)acetone | |
| CA2514945A1 (en) | Methods for producing quinazoline alkaloids | |
| JPWO2005058859A1 (en) | Process for producing 3- (4-tetrahydropyranyl) -3-oxopropanoic acid alkyl compound and 4-acyltetrahydropyran | |
| US6916792B2 (en) | Process for preparing erythromycin compound | |
| KR20000018793A (en) | Method for manufacturing 1,2-benzisothiazolones-3-one | |
| CN118239969A (en) | Preparation method of benzo-1, 4-dioxane-6-boric acid | |
| CN115677619A (en) | Method for preparing sulfonate by oxidizing imidosulfonate with sodium hypochlorite | |
| KR101315751B1 (en) | New method for producing Loperamide oxide monohydrate | |
| Talaty et al. | Adducts of α-Lactams and a Dihydro-1, 3-Oxazine: A Convenient Synthesis of α-Amino Ketones | |
| US20040204573A1 (en) | Process for preparation of erythromycin compounds | |
| ZA202304623B (en) | Water-phase synthesis method of tetrabromo phenol tetrahalogenated suofonephthalein |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 201080054252.2 Country of ref document: CN |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 10773387 Country of ref document: EP Kind code of ref document: A1 |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2010773387 Country of ref document: EP |
|
| ENP | Entry into the national phase |
Ref document number: 2780027 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1125/MUMNP/2012 Country of ref document: IN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2010309541 Country of ref document: AU |
|
| ENP | Entry into the national phase |
Ref document number: 2010309541 Country of ref document: AU Date of ref document: 20101022 Kind code of ref document: A |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 13522455 Country of ref document: US |