WO2010109476A2 - Improved process for the preparation of prostaglandins and analogues thereof - Google Patents
Improved process for the preparation of prostaglandins and analogues thereof Download PDFInfo
- Publication number
- WO2010109476A2 WO2010109476A2 PCT/IN2010/000026 IN2010000026W WO2010109476A2 WO 2010109476 A2 WO2010109476 A2 WO 2010109476A2 IN 2010000026 W IN2010000026 W IN 2010000026W WO 2010109476 A2 WO2010109476 A2 WO 2010109476A2
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- formula
- compound
- solvent
- process according
- base
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 C[C@@](C[C@@](*)CCc1ccccc1)[C@@]([C@](*)C1)[C@](CC=CCCCC(O)=O)[C@]1O Chemical compound C[C@@](C[C@@](*)CCc1ccccc1)[C@@]([C@](*)C1)[C@](CC=CCCCC(O)=O)[C@]1O 0.000 description 3
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C405/00—Compounds containing a five-membered ring having two side-chains in ortho position to each other, and having oxygen atoms directly attached to the ring in ortho position to one of the side-chains, one side-chain containing, not directly attached to the ring, a carbon atom having three bonds to hetero atoms with at the most one bond to halogen, and the other side-chain having oxygen atoms attached in gamma-position to the ring, e.g. prostaglandins ; Analogues or derivatives thereof
Definitions
- the present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues, particularly PGF 2 ⁇ derivatives.
- Prostaglandin analogs represent the most potent therapeutic agents in the clinical management of ocular hypertension and glaucoma.
- Prostaglandins are generally characterized by the substituents on the cyclopentyl ring.
- the PGF 2 Q prostaglandins and prostaglandin analogues generally have two hydroxyl groups in a cis configuration relative to the cyclopentane ring, and two side chains in a trans configuration relative to each other, each side chain having one double bond.
- Analogues of PGF 2 ⁇ can have a different number of double bonds in the side chains, and the substituents along the side chains may vary. Additionally, in some PGF 20 analogues, the side chain carboxylic acid group may be esterified.
- Prostaglandin analogues based on PGF 2 ⁇ for use in the treatment of glaucoma and ocular hypertension are described in, for example, European patent number 0 364 417 B1.
- the procedures for the synthesis of PGF 20 analogues described therein start from an advanced- stage intermediate, 16-phenyl-17,18,19,20-trinor PGF 2 ⁇ , or the tetranor homologue thereof
- the isopropyl ester of PGF 2 ⁇ commonly called as Latanoprost [13,14-dihydro-15(R)-17- phenyl-18,19,20-trinor-PGF 2 ⁇ -isopropyl] has been shown to have significantly greater hypotensive potency, useful in the reduction of elevated intra-ocular pressure in patients with open angle glaucoma and ocular hypertension.
- prostaglandin analogs used to reduce elevated intraocular pressure apart from latanoprost include travoprost (Travatan) and bimatoprost (Lumigan) etc.
- an object of the present invention is to provide an alternative process for the synthesis of PGF 2 ⁇ and analogues thereof, particularly Latanoprost and Bimatoprost.
- the main object of the present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
- Another object of the present invention is related to the synthesis of PGF 20 analogues such as latanoprost and bimatoprost.
- Yet another object of the present invention is related to the synthesis of a bis[tert-butyl dimethylsilyloxy] protected carboxylic acid intermediate for the preparation of Latanoprost and
- Yet another object of the present invention is to prepare Latanoprost and Bimatoprost by the deprotection of bis[tert-butyl dimethylsilyloxy] protected carboxylic acid intermediate.
- the present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
- prostaglandin derivatives of the present invention are represented by formula IA
- R' represents COOH, ester groups such as COOAIk wherein AIk independently represent methyl, ethyl, propyl, isopropyl and the like, amides such as CONHAIk wherein AIk independently represent methyl, ethyl, propyl, isopropyl and the like.
- R" represents optionally substituted aryl such as phenyl, optionally substituted arylalkyl such as benzyl, optionally substituted aryloxy such as phenoxy wherein the said aryl, arylalkyl or aryloxy groups may be substituted with one or more groups independently selected from hydroxy, halides such as F, Cl, Br or I, lower alkyl such as methyl, ethyl or alkyl halides such as CH 2 F, CHF 2 , CF 3 and the like.
- the dotted bond ( ) represents an optional bond.
- the present invention provides a process for the preparation of Latanoprost comprising the steps of; (a) oxidizing protected primary alcohol ( LTN-A) to aldehyde (LTN-B), (b) treating aldehyde (LTN-B) with triphenyl-4(phenyl-2- oxobutyl)phosphonium bromide (SC-1) to obtain enone (LTN - C), (c) treating enone (LTN-C) with (-) DIP chloride to obtain hydroxyl analogue (LTN-D), (d) hydrogenating the hydroxyl analogue (LTN-D) in the presence of a catalyst to obtain LTN-E, (e) deprotecting LTN-E to obtain the lactone (LTN-F), (f) treating lactone (LTN-F) with tert-butyldimethylsilyl chloride to obtain diprotected lactone (LTNG), (g) reducing di-protected lactone (LTN-A)
- the present invention provides a process for the preparation of Bimatoprost comprising the steps of; (a) oxidizing protected primary alcohol to aldehyde (BPT -1), (b) treating aldehyde (BPT-1) with triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) to obtain enone (BPT-2), (c) treating enone (BPT-2) with (-) DIP chloride to obtain hydroxyl analogue (BPT-3), (d) deprotecting (BPT-3) to obtain lactone derivative (BPT- 4), (e) treating lactone derivative (BPT-4) with tert-Butyl Dimethyl silyl chloride to obtain diprotected lactone derivative (BPT-5), (f) reducing di-protected lactone (BPT -5) into di- protected lactol (BPT-6), (g) treating di-protected lactol (BPT -6) with 4-carboxy butyl trip
- Fig 1 shows an X-ray Diffraction diagram of Bimatoprost. DETAILED DESCRIPTION OF THE INVENTION
- the present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
- the present invention relates to an improved process for the preparation of prostaglandins and prostaglandin analogues.
- the present invention relates to the synthesis of PGF 20 analogues such as latanoprost and bimatoprost.
- the present invention encompasses an improved process for the preparation of latanoprost as summarized in scheme 1.
- the present invention provides a process for the preparation of latanoprost comprising the steps of; a) oxidizing protected primary alcohol ( LTN-A) to aldehyde (LTN-B), b) treating aldehyde (LTN-B) with triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) in the presence of solvent to obtain enone (LTN - C), c) treating enone (LTN-C) with (-) DIP chloride in the presence of a solvent to obtain hydroxyl analogue (LTN-D), d) hydrogenating the hydroxyl analogue (LTN-D) in the presence of a catalyst, base and solvent to obtain LTN-E, e) deprotecting LTN-E in the presence of a base and solvent to obtain the lactone (LTN-A) to aldehyde (LTN-B), b) treating aldehyde (LTN-B) with triphenyl-4(
- the present invention encompasses an improved process for the preparation of Bimatoprost as summarized in scheme 2.
- the present invention provides a process for the preparation of bimatoprost comprising the steps of: a) oxidizing protected primary alcohol to aldehyde (BPT -1), b) treating aldehyde (BPT-1) with triphenyl ⁇ 4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) to obtain enone (BPT - 2), c) treating enone (BPT-2) with (-) DIP chloride in the presence of solvent to obtain hydroxyl analogue (BPT-3), d) deprotecting (BPT-3) in the presence of a base and a solvent to obtain lactone derivative (BPT-4), e) treating lactone derivative (BPT-4) with tert-Butyl Dimethyl silyl chloride in the presence of a solvent to obtain diprotected lactone derivative (BPT-5), f) reducing di-protected lactone (BPT -5) in the presence of reducing agent, solvent into di-protected
- the oxidizing agents used in the oxidation of the protected primary alcohol to aldehyde are selected from a combination of dimethyl sulfoxide with dicyclohexylcarbodiimide.
- the oxidation reaction is carried out in the presence of an acid selected from phosphoric acid and in a solvent selected from dimethoxy ethane or toluene or mixtures thereof.
- reaction of aldehyde with triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) is carried out in the presence of solvent selected from dichloromethane or toluene or mixtures thereof.
- the solvent used in reaction of enone with (-) DIP chloride is selected from tetrahydrofuran, dichloromethane or toluene or mixtures thereof.
- the reduction of double bond is carried out using a catalyst selected from palladium on carbon, platinum or nickel, preferably palladium on carbon.
- Reduction reaction is carried out in the presence of base and solvent, base is selected from sodium hydroxide or potassium hydroxide, solvent is selected from ethanol, methanol, isopropanol, n-butanol or mixtures thereof.
- the aroyl deprotection is carried out in presence of a base and solvent selected from potassium carbonate, sodium carbonate, cesium carbonate, potassium bicarbonate, sodium bicarbonate.
- the solvent is selected from ethanol, methanol, isopropanol, n-butanol or mixtures thereof.
- the diol protection is carried out using tert-butyldimethylsilyl chloride in the presence of a base and solvent.
- the base is selected from imidazole, triethylamine, diisopropylethylamine or tributylamine.
- Solvent group is from polar aprotic solvents such as tetrahydrofuran or dimethylsulfoxide, or chlorinated solvents such as dichloromethane, chloroform, 1,2- dichloroethane.
- lactone to lactol is carried out using diisobutyl aluminium hydride in the presence of solvent selected from toluene, tetrahydrofuran, diethyl ether or dichloromethane, or mixtures thereof.
- the Wittig reaction of di-protected lactol with 4-carboxy butyl triphenyl phosphonium bromide in the presence of base is selected from butyl lithium, sodium amide, sodium hydride, sodium methoxide, sodium ethoxide, potassium tert-butoxide or potassium ethoxide, preferably potassium tert-butoxide is used.
- the solvent used for the Wittig reaction is selected from tetrahydrofuran, toluene, dichloromethane or mixtures thereof.
- the esterification of the carboxylic acid derivative (LTN-I) is carried out with isopropyl bromide to get the ester derivative (LTN-J) in the presence of a base selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate, sodium bicarbonate, preferably cesium carbonate.
- a base selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate, sodium bicarbonate, preferably cesium carbonate.
- the solvent for the esterification reaction is selected from dimethyl formamide, dichloromethane, isopropanol, acetonitrile, tetrahydrofuran, acetone, methyl ethyl ketone or mixtures thereof.
- the esterification of carboxylic acid derivative (BPT-7) is carried out with methyl iodide in the presence of base and solvent to get the ester derivative (BPT-8).
- the base is selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate , sodium bicarbonate, preferably cesium carbonate.
- the solvent for the esterification reaction is selected from dimethyl formamide, dichloromethane, isopropanol, acetonitrile, tetrahydrofuran, acetone, methyl ethyl ketone or mixtures thereof.
- the amidification of the ester (BPT-8) is carried out by using ethylamine gas in presence of solvent to get the diprotected Bimatoprost (BPT-9).
- Solvent is selected from methanol, ethanol or mixtures thereof;
- the final hydrolysis and the silyl group deprotection (for both LTN-J and BPT-9) is carried out in the presence of an acid selected from aqueous hydrochloric acid or aqueous sulfuric acid in the presence of a solvent selected from isopropanol, isobutanol, n-propanol, n-butanol, ethanol or methanol mixtures thereof.
- the present invention provides a process for the preparation of the compound of formula- Y
- the acid used herein for deprotection is selected from aq. hydrochloric acid or aq. sulfuric acid.
- the solvents used herein are selected from methanol, ethanol, isopropyl alcohol, n-butanol or n-propanol.
- the reaction is performed at 25°C to 50 0 C, more preferably at 25°C to 30°C.
- the present invention provides a process for the purification of Bimatoprost comprising the steps of; a) dissolving Bimatoprost in a solvent, b) adding of an antisolvent, and c) isolating Bimatoprost.
- the solvent is selected from chlorinated solvents like dichloromethane, chloroform, 1,2-dichloroethane, carbon tetrachloride, or mixtures thereof.
- the resulting solution is passed through a micron filter, like hyflow bed or silica gel bed.
- the anti-solvent is selected from diisopropyl ether and diethyl ether, preferably diisopropyl ether.
- the product may be isolated by the conventional techniques of filtration or centrifugation.
- the present invention provides a process for the preparation of triphenyl-4(phenyl-2-oxobutyl) phosphonium bromide (SC-1) as shown below
- the present invention provides a process for the preparation of triphenyl-4(phenyl-2-oxobutyl) phosphonium bromide (SC-1), comprising the steps of; a) treating benzyl acetone with bromine to obtain the bromo derivative. b) treating the bromo derivative with triphenyl phosphine in a solvent to get triphenyl- 4(phenyl-2- oxo butyl)phosphonium bromide.
- SC-1 triphenyl-4(phenyl-2-oxobutyl) phosphonium bromide
- benzyl acetone is reacted with alcoholic bromine solution in the presence of a solvent selected from methanol, ethanol, isopropanol, n-propanol or mixtures thereof.
- DIBAL reaction is either incomplete or forms more impurities when both the hydroxyl groups are unprotected or even one hydroxyl group is protected.
- DIBAL reaction proceeds smoothly (with no formation of impurities) within 60-90 minutes with better yield and quality of product, vi.
- Step-1 Preparation of Triphenyl-4(phenyl-2-oxo butyl) phosphonium bromide A) In a one litre round bottom flask fitted with an overhead stirrer, thermo pocket and dropping funnel was charged methanol (270ml) and benzyl acetone (100g) at 20 to 30 0 C.
- Bromine solution (118.8 g bromine dissolved in 270 ml methanol) was added during 60 to 90 minutes maintaining internal temperature at 7 to10 ° C.
- the reaction mass was maintained at same temperature for 2 to 4 hours and checked HPLC for completion of reaction.
- D. M. water (674 ml) was added at 7 to 10 0 C.
- the temperature was raised to 20 to 30° C and maintained for 12 hours at the same temperature.
- the organic layer was separated.
- the upper aqueous layer was extracted with dichloromethane (270ml) and combined with the organic layer. Dichloromethane was distilled out completely under reduced pressure to obtain viscous residue (140-150 g) which was dissolved in 300 ml toluene.
- Step-2 Preparation of (1S,5R,6R,7R)-6-Oxo-5-phenyl-1E-pentenyl)-7-
- Dimethoxy ethane (60ml) was charged into Corey lactone (10 g) at 25 to 30°c in a 250 ml four neck round bottom flask equipped with mechanical stirrer and dropping funnel. The suspension was stirred for 10 minutes, charged dicyclohexyl carbodiimide (23.3) and dimethyl sulfoxide (DMSO) (17.75 g) at 25 to 30 0 C. The reaction mass was cooled to 20 to 22 0 C and phosphoric acid was added (1.65g). The reaction mixture was maintained at 20 to 22° C for about 20 to 30 minutes. The temperature was raised to 25 to 30 0 C. After completion of reaction, the unwanted dicyclohexyl urea was filtered and washed with dimethoxy ethane (20 ml). The filtrate contains Corey aldehyde.
- reaction mass was cooled to 18 to 2O 0 C and dilute HCI (2.5 ml cone. HCL diluted in 17.5 ml water) was added to it.
- the reaction mass was stirred for 15 to 20 minutes, filtered, and the organic layer was separated. The organic layer was washed twice with water (2X50 ml). The solvent was distilled out completely under vacuum and the residue was charged with ethanol
- Step-3 Preparation of (1S, 5R, 6R, 7R)-6-[ (3S)-3- hydroxyl-5-phenyl-1E-pentenyl)-7-[ (4- phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one.
- Step-3A Preparation of ("IS, 5R, 6R, 7R)-6-[ (3S)-3- hydroxyl-5-phenyl-1E-pentenyl)-7-[ (4-phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one.
- Step 1 Preparation of 1S,5R,6R,7R)-6-[(3R)-3-Hydroxy-5-phenyl-1pentyl)-7-(4- phenyl benzoyl)oxy ⁇ -2-oxabicyclo[3.3.0] octan-3-one
- Step 2 Preparation of 1S,5R,6R,7R)-6-[(3R)-3-Hydroxy-5-phenyl-1-pentyl)-7-(S)- hydroxy-2-oxabicyclo[3.3.0] octan-3-one
- Step 3 Preparation of 1S,5R,6R,7R)-6-[(3S)dimethyltert-butylsilyloxy-5-phenyl-1- pentyl)(7S)dimethyltert-butylsilyloxy-2-oxabicyclo[3.3.0]octan-3-one
- Step 4 Preparation of 1S,5R,6R,7R)-6-[(3R)dimethyl tert-butylsilyloxy-5-phenyl-1-pentyl) (7S) dimethyl t.butyl silyloxy -2-oxabicyclo[3.3.0] octan-3-ol
- Toluene (320 ml) was charged into the compound isolated in step 3 of example-1 (32 g) in a 500 ml round bottom flask equipped with overhead stirrer, thermo pocket and dropping funnel at 25 to 30°.
- the solution was cooled to -70 0 C and slowly added Diisobutyl aluminum hydride (1 M solution in toluene, 130 ml) maintaining the temperature at -70 to -80 0 C.
- the reaction mixture was stirred at same temperature for 90 min and progress of reaction was monitored by TLC.
- methanol (130 ml) was charged at -70 to -80°C. Raised the temperature from 25 to 30 0 C and maintained for 30 minutes.
- the unwanted salt was filtered and washed with methanol (30 ml).
- the filtrate was passed through silica column and solvent was distilled out completely under vacuum to obtained above said product as viscous oil (33 g) (HPLC purity - 93%)
- Step 5 Preparation of (Z)-7-(1R,2R,3R,5S)-5-hydroxy-2-[(3R)dimethyltert-butyl silyloxy- 5-(phenyl-1 -pentyl)-3-(dimethyl tert-butylsilyloxy)cyclopentyl-5-hetp-enoic acid.
- Tetrahydrofuran was distilled out completely under vacuum below 30 0 C.
- Water (100 ml) was charged and stirred for 30 minutes at 25 to 3O 0 C.
- the unwanted salt was filtered and washed with water (10 ml).
- citric acid solution was added to adjust pH to 4.0.
- Ethyl acetate (250 ml) was added and stirred for 20 minutes.
- the organic layer was separated and the aqueous layer was extracted with ethyl acetate (250 ml).
- the combined ethyl acetate layers were washed with brine solution and distilled out completely under vacuum at 40-45°C.
- the crude product thus obtained was purified by column chromatography to get 7-8 grams title compound.
- Step 6 Preparation of lsopropyl (Z)-7-(1R I 2R,3R,5S)-5-hydroxy-2-[(3R)dimethyl tert- butylsilyloxy-5-(phenyl-1-pentyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5-hetpanoate
- Dimethyl formamide (100 ml) and compound of step 5 of example 1 (10 g) were taken in a 250 ml round bottom flask at 25 to 30°C.
- Cesium carbonate (7.8 g) was charged under stirring at 25 to 30°C and stirred for 15 minutes.
- step 3 To a solution of product obtained in step 3 (10 g) in methanol (100 ml) was charged potassium carbonate (1.86 g) at 20-30°C and stirred at 20 to 30°C for 5 to 7 hours. After the completion of the reaction, methanol was distilled out completely under vacuum at 40 to 45°C. 1N Hydrochloric acid (120 ml) and ethyl acetate (100 ml) were charged and stirred for 30 minutes at 20 to 30°C. The organic layer was separated and the aqueous layer was extracted with ethyl acetate twice. The combined ethyl acetate layers were washed with brine solution and ethyl acetate was distilled out completely under vacuum at 40 to 45°C. The residue thus obtained was separated by column chromatography to obtain 5.4 to 6.0 g title compound as viscous oil.
- Step 2 Preparation of 1S,5R,6R,7R)-6-[(3S)dimethyltert-butylsilyloxy-5-phenyl-1- pentenyl) (7S)dimethyltert-butyl silyloxy-2-oxabicyclo[3.3.0]octan-3-one
- tetrahydrofuran (170 ml)
- 1S,5R,6R,7R-6-[(3R)-3-hydroxy-5-phenyl-1-pentenyl)-7-(S)- hydroxy-2-oxabicyclo [3.3.0] octan-3-one (16 g) at 25 to 30°C.
- the reaction mixture was stirred for 10 minutes followed by the addition of imidazole (21.6 g) and tert-Butyld imethylsilyl chloride (33.6 g) at 25 to 30°C. The temperature was raised to 65 to 70 0 C and maintained for 60 to 90 minutes until the completion of the reaction. The reaction mass was cooled to 40 to 45°C and tetrahydrofuran was distilled out completely under vacuum. Dichloromethane (160 ml) was charged at 25 to 30 0 C and stirred for 10 minutes to get the clear solution. The reaction mass was washed with 10% citric acid (50 ml). The solvent was distilled out completely under vacuum at 40 to 45°C.
- Methyl tert-Butyl ether (MTBE) 50 ml was charged to the residue and stirred for 15 minutes to get clear solution. MTBE was distilled out completely under vacuum at 40 to 45°C to get 27-29 g of title compound as a white residue.
- Step 3 Preparation of 1S,5R,6R,7R)-6-[(3R)dimethyltert-butylsilyloxy-5-phenyl-1- pentenyl) (7S) dimethyltert-butyl silyloxy-2-oxabicyclo[3.3.0] octan-3-ol
- Toluene (260 ml) was charged into the compound isolated in step 2 of example 2 (25 g) in a 500 ml round bottom flask equipped with overhead stirrer, thermo pocket and dropping funnel at 25 to 3O 0 C.
- the reaction mixture was cooled to -70 0 C and slowly added Diisobutyl aluminum hydride (122 ml) (20% solution in toluene) maintaining the temperature at -70 to - 80°C. Stirred at same temperature for 45 minutes and the progress of reaction was monitored was by TLC. Methanol (105 ml) was charged at -70 to -80°C after completion of the reaction. The temperature was raised to 25 to 3O 0 C and maintained for 30 minutes. The unwanted salt was filtered and and washed with methanol (50 ml). The filtrate was passed through silica column and solvent was distilled out completely under vacuum to obtain the above said product as viscous oil (23 g).
- Step 4 Preparation of (Z)- 7-(1R,2R,3R,5S) -5-hydroxy -2- [(3R) dimethyl tbutyl silyloxy - 5-(phenyl-1-pentenyl)-3-( dimethyl t.butyl silyloxy) cyclopentyl -5-hetp-enoic acid.
- (4-Carboxybutyl)triphenylphosphonium bromide (151.5 g) was suspended in tetrahydrofuran (604 ml) at 25 to 30 0 C.
- reaction mixture was cooled under stirring to 15 to 20 0 C and lot wise potassium tertiary butoxide (76.8 g) was added at 15 to 20° C and maintained for 30 minutes. Cooled slowly to -10 to -15° C and a solution of compound obtained in step 3 of example 2 (22.8 g in 69 ml tetrahydrofuran) was added slowly at -10 to -15°C. Maintained at - 10 to -15°C for 2 hours until TLC shows the reaction was complete. The tetrahydrofuran was distilled out completely under vacuum below 30°C. Water (228 ml) was charged and stirred for 30 minutes at 25 to 30 0 C. The unwanted salt was filtered and washed with water (25 ml).
- Step 5 Preparation of Methyl (Z)-7-(1R,2R,3R,5S)-5-hydroxy-2-[(3R)dimethyl tert- butylsilyloxy-5-(phenyl-1-pentenyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5- hetpanoate
- Step 6 Preparation of (Z)-7-(1R,2R,3R I 5S)-5-hydroxy-2-[(3R)dimethyltert-butyl silyloxy- 5-(phenyl-1-pentenyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5-N-ethyl heptenamide (protected Bimatoprost)
- thermo pocket and stopper was charged methanol (20 ml) and the compound obtained in step 6 of example 2 at 25 to 30 0 C.
- the resulting mixture was cooled to 15 to 20 0 C and slowly added 2N HCI (5 ml) at 15 to 20 0 C.
- the reaction mixture was maintained under stirring at 15 to 20 0 C for 3 to 5 hours till reaction is completed by TLC.
- Water 40 ml was added and cooled to 0 to 10 0 C, the pH was adjusted to 6.5 to 7.0 with 5% sodium carbonate solution at 0 to 10 0 C.
- Ethyl acetate (40 ml) was charged and stirred for 15 minutes.
- the crude Bimatoprost (1.2 g) was dissolved in dichloromethane (15 ml) at 20-25°C to get clear solution. The reaction mass was filtered through micron filter to make particle free. Charged filtrate to RB flask and slowly added diisopropyl ether (90 ml) over a period of 30-45 minutes. The product was filtered and washed with diisopropyl ether to obtain 0.8-0.9 g pure Bimatoprost.
- Step 8A Purification of Bimatoprost
- step 7 The crude product obtained in step 7 (1.2 g) was dissolved in dichloromethane (15 ml) to get clear solution at 25 to 300 C. slowly added diisopropyl ether (90 ml) over a periodof 30 to 45 minutes. Stirred for one hour at 25 to 300 C, filtered and washed with diisopropyl ether to obtain 0.8 to 0.9 g pure bimatoprost
- diethyl ether can also be used instead of diisopropyl ether.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Abstract
The present invention relates to an improved process for the preparation of prostaglandin and prostaglandin analogues, particularly PGF2a derivatives.
Description
IMPROVED PROCESS FOR THE PREPARATION OF PROSTAGLANDINS AND
ANALOGUES THEREOF
FIELD OF THE INVENTION
The present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues, particularly PGF2α derivatives.
BACKGROUND OF THE INVENTION
Prostaglandin analogs represent the most potent therapeutic agents in the clinical management of ocular hypertension and glaucoma.
Prostaglandins are generally characterized by the substituents on the cyclopentyl ring. The PGF2Q prostaglandins and prostaglandin analogues generally have two hydroxyl groups in a cis configuration relative to the cyclopentane ring, and two side chains in a trans configuration relative to each other, each side chain having one double bond. Analogues of PGF2α can have a different number of double bonds in the side chains, and the substituents along the side chains may vary. Additionally, in some PGF20 analogues, the side chain carboxylic acid group may be esterified.
Prostaglandin analogues based on PGF2α for use in the treatment of glaucoma and ocular hypertension are described in, for example, European patent number 0 364 417 B1. The procedures for the synthesis of PGF20 analogues described therein start from an advanced- stage intermediate, 16-phenyl-17,18,19,20-trinor PGF2α, or the tetranor homologue thereof The isopropyl ester of PGF2α commonly called as Latanoprost [13,14-dihydro-15(R)-17- phenyl-18,19,20-trinor-PGF2α-isopropyl], has been shown to have significantly greater hypotensive potency, useful in the reduction of elevated intra-ocular pressure in patients with open angle glaucoma and ocular hypertension.
The other marketed prostaglandin analogs used to reduce elevated intraocular pressure apart from latanoprost (Xalatan) include travoprost (Travatan) and bimatoprost (Lumigan) etc.

Latanoprost Bimatoprost

Travoprost
A variety of methods of synthesizing PGF2α analogs are known and are disclosed in for e.g. US 7166730, US 6403649, EP 1721894, US 7157590, US 5698733, US7163959, US 5359095, US 6927300, WO 02/096868, WO 02/090324, Chem. Rev.(1993, vol. 93, pages 1533-1564), Chinese journal of medicinal chemistry (1998, vol. 36, pages 213-217), B. Resul et al., Journal of Medicinal Chemistry., 1993, 36, 243)
A typical synthesis of prostaglandin derivative is disclosed in WO 93/00329 which is summarized below:

Use of silyl protecting groups for the protection of dihydroxy groups is disclosed US 7268239.

It is well known that by using silylating agents for the protection of hydroxyl groups, the reactions are clean and yields are quantitative.
The processes disclosed herein above have the below disadvantages
i. Lot of impurities were formed when unprotected hydroxy I groups are used and it requires the purification of the intermediates at almost every stage of synthesis which leads to very low yields and time consuming ii. Conventional protecting agents for hydroxyl groups again resulted in some difficulties in the subsequent steps iii. The use of separate protecting groups for the two hydroxyl groups leads to an additional step with different reagent iv. Deprotection may lead to problems with different protecting groups, wherein both the hydroxyl groups may not be deprotected at the same time.
In view of the processes disclosed herein above and the problems associated therewith, it is an object of the present invention is to provide an alternative process for the synthesis of PGF2α and analogues thereof, particularly Latanoprost and Bimatoprost.
OBJECT OF THE INVENTION
The main object of the present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
Another object of the present invention is related to the synthesis of PGF20 analogues such as latanoprost and bimatoprost.
Yet another object of the present invention is related to the synthesis of a bis[tert-butyl dimethylsilyloxy] protected carboxylic acid intermediate for the preparation of Latanoprost and
Bimatoprost.
Yet another object of the present invention is to prepare Latanoprost and Bimatoprost by the deprotection of bis[tert-butyl dimethylsilyloxy] protected carboxylic acid intermediate.
SUMMARY OF THE INVENTION
The present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
The prostaglandin derivatives of the present invention are represented by formula IA

Formula IA wherein
R' represents COOH, ester groups such as COOAIk wherein AIk independently represent methyl, ethyl, propyl, isopropyl and the like, amides such as CONHAIk wherein AIk independently represent methyl, ethyl, propyl, isopropyl and the like.
R" represents optionally substituted aryl such as phenyl, optionally substituted arylalkyl such as benzyl, optionally substituted aryloxy such as phenoxy wherein the said aryl, arylalkyl or aryloxy groups may be substituted with one or more groups independently selected from hydroxy, halides such as F, Cl, Br or I, lower alkyl such as methyl, ethyl or alkyl halides such as CH2F, CHF2, CF3 and the like. The dotted bond ( ) represents an optional bond.
According to one aspect, the present invention provides a process for the preparation of Latanoprost comprising the steps of; (a) oxidizing protected primary alcohol ( LTN-A) to aldehyde (LTN-B), (b) treating aldehyde (LTN-B) with triphenyl-4(phenyl-2- oxobutyl)phosphonium bromide (SC-1) to obtain enone (LTN - C), (c) treating enone (LTN-C) with (-) DIP chloride to obtain hydroxyl analogue (LTN-D), (d) hydrogenating the hydroxyl analogue (LTN-D) in the presence of a catalyst to obtain LTN-E, (e) deprotecting LTN-E to obtain the lactone (LTN-F), (f) treating lactone (LTN-F) with tert-butyldimethylsilyl chloride to obtain diprotected lactone (LTNG), (g) reducing di-protected lactone (LTN-G) to obtain di- protected lactol (LTN-H), (h) reacting (LTN-H) with 4-carboxy butyl triphenyl phosphonium bromide to obtain diprotected lactol (LTN-I), (i) treating diprotected lactol (LTN-I) with isopropyl bromide to obtain an ester (LTN-J), (j) deprotecting the ester derivative (LTN-J) to obtain latanoprost.
According to another aspect, the present invention provides a process for the preparation of Bimatoprost comprising the steps of; (a) oxidizing protected primary alcohol to aldehyde (BPT -1), (b) treating aldehyde (BPT-1) with triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) to obtain enone (BPT-2), (c) treating enone (BPT-2) with (-) DIP chloride to obtain hydroxyl analogue (BPT-3), (d) deprotecting (BPT-3) to obtain lactone derivative (BPT- 4), (e) treating lactone derivative (BPT-4) with tert-Butyl Dimethyl silyl chloride to obtain diprotected lactone derivative (BPT-5), (f) reducing di-protected lactone (BPT -5) into di-
protected lactol (BPT-6), (g) treating di-protected lactol (BPT -6) with 4-carboxy butyl triphenyl phosphonium bromideto obtain the diprotected acid derivative (BPT-7), (h) esterifying the diprotected acid derivative (BPT-7) to obtain ester derivative (BPT-8), (i) converting the ester derivative (BPT-8) into corresponding amide derivative (BPT-9), (j) deprotecting the amide derivative (BPT-9) to obtain bimatoprost.
BRIEF DESCRIPTION OF THE FIGURES
Fig 1 shows an X-ray Diffraction diagram of Bimatoprost. DETAILED DESCRIPTION OF THE INVENTION
The present invention is related to an improved process for the preparation of prostaglandin and prostaglandin analogues.
In one embodiment, the present invention relates to an improved process for the preparation of prostaglandins and prostaglandin analogues. In another embodiment, the present invention relates to the synthesis of PGF20 analogues such as latanoprost and bimatoprost.
In yet another embodiment, the present invention encompasses an improved process for the preparation of latanoprost as summarized in scheme 1.
Scheme 1

F), f) treating lactone (LTN-F) with tert-butyldimethylsilyl chloride in the presence of an organic base and solvent to obtain diprotected lactone (LTN-G), g) reducing di-protected lactone (LTN-G) in the presence of solvent to obtain di- protected lactol (LTN-H), h) reacting (LTN-H) with 4-carboxy butyl triphenyl phosphonium bromide in the presence of a base and a solvent to obtain diprotected lactol (LTN-I), i) esterifying the diprotected lactol (LTN-I) with isopropyl bromide in the presence of base to obtain an ester (LTN-J), and j) deprotecting the ester derivative (LTN-J) to obtain latanoprost.
In yet another embodiment the present invention encompasses an improved process for the preparation of Bimatoprost as summarized in scheme 2.
Scheme 2

In yet another embodiment the present invention provides a process for the preparation of bimatoprost comprising the steps of: a) oxidizing protected primary alcohol to aldehyde (BPT -1), b) treating aldehyde (BPT-1) with triphenyl~4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) to obtain enone (BPT - 2), c) treating enone (BPT-2) with (-) DIP chloride in the presence of solvent to obtain hydroxyl analogue (BPT-3), d) deprotecting (BPT-3) in the presence of a base and a solvent to obtain lactone derivative (BPT-4), e) treating lactone derivative (BPT-4) with tert-Butyl Dimethyl silyl chloride in the presence of a solvent to obtain diprotected lactone derivative (BPT-5), f) reducing di-protected lactone (BPT -5) in the presence of reducing agent, solvent into di-protected lactol (BPT-6),
g) treating di-protected lactol (BPT -6) with 4-carboxy butyl triphenyl phosphonium bromide in the presence of a base, solvent to obtain the diprotected acid derivative (BPT-7), h) esterifying the diprotected acid derivative (BPT-7) in the presence of base to obtain ester derivative (BPT-8), i) reacting the ester derivative (BPT-8) into corresponding amide derivative (BPT-9), and j) deprotecting the amide derivative (BPT-9) to obtain bimatoprost.
According to the present invention, the oxidizing agents used in the oxidation of the protected primary alcohol to aldehyde are selected from a combination of dimethyl sulfoxide with dicyclohexylcarbodiimide. The oxidation reaction is carried out in the presence of an acid selected from phosphoric acid and in a solvent selected from dimethoxy ethane or toluene or mixtures thereof.
The reaction of aldehyde with triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide (SC-1) is carried out in the presence of solvent selected from dichloromethane or toluene or mixtures thereof.
The solvent used in reaction of enone with (-) DIP chloride [(-)-Diisopinocampheyl chloroborane] is selected from tetrahydrofuran, dichloromethane or toluene or mixtures thereof.
The reduction of double bond is carried out using a catalyst selected from palladium on carbon, platinum or nickel, preferably palladium on carbon. Reduction reaction is carried out in the presence of base and solvent, base is selected from sodium hydroxide or potassium hydroxide, solvent is selected from ethanol, methanol, isopropanol, n-butanol or mixtures thereof.
The aroyl deprotection is carried out in presence of a base and solvent selected from potassium carbonate, sodium carbonate, cesium carbonate, potassium bicarbonate, sodium bicarbonate. The solvent is selected from ethanol, methanol, isopropanol, n-butanol or mixtures thereof.
The diol protection is carried out using tert-butyldimethylsilyl chloride in the presence of a base and solvent. The base is selected from imidazole, triethylamine, diisopropylethylamine or
tributylamine. Solvent group is from polar aprotic solvents such as tetrahydrofuran or dimethylsulfoxide, or chlorinated solvents such as dichloromethane, chloroform, 1,2- dichloroethane.
The reduction of lactone to lactol is carried out using diisobutyl aluminium hydride in the presence of solvent selected from toluene, tetrahydrofuran, diethyl ether or dichloromethane, or mixtures thereof.
The Wittig reaction of di-protected lactol with 4-carboxy butyl triphenyl phosphonium bromide in the presence of base. The base is selected from butyl lithium, sodium amide, sodium hydride, sodium methoxide, sodium ethoxide, potassium tert-butoxide or potassium ethoxide, preferably potassium tert-butoxide is used. The solvent used for the Wittig reaction is selected from tetrahydrofuran, toluene, dichloromethane or mixtures thereof.
The esterification of the carboxylic acid derivative (LTN-I) is carried out with isopropyl bromide to get the ester derivative (LTN-J) in the presence of a base selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate, sodium bicarbonate, preferably cesium carbonate. The solvent for the esterification reaction is selected from dimethyl formamide, dichloromethane, isopropanol, acetonitrile, tetrahydrofuran, acetone, methyl ethyl ketone or mixtures thereof.
The esterification of carboxylic acid derivative (BPT-7) is carried out with methyl iodide in the presence of base and solvent to get the ester derivative (BPT-8). The base is selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate , sodium bicarbonate, preferably cesium carbonate. The solvent for the esterification reaction is selected from dimethyl formamide, dichloromethane, isopropanol, acetonitrile, tetrahydrofuran, acetone, methyl ethyl ketone or mixtures thereof.
The amidification of the ester (BPT-8) is carried out by using ethylamine gas in presence of solvent to get the diprotected Bimatoprost (BPT-9). Solvent is selected from methanol, ethanol or mixtures thereof;
The final hydrolysis and the silyl group deprotection (for both LTN-J and BPT-9) is carried out in the presence of an acid selected from aqueous hydrochloric acid or aqueous sulfuric acid in
the presence of a solvent selected from isopropanol, isobutanol, n-propanol, n-butanol, ethanol or methanol mixtures thereof.
In yet another embodiment, the present invention provides a process for the preparation of the compound of formula- Y

Formula-Y wherein P is either O1Pr or NHEt, by deprotection of the compound of formula-X

Formula-X in the presence of an acid with or without solvent.
According to the present invention the acid used herein for deprotection is selected from aq. hydrochloric acid or aq. sulfuric acid. The solvents used herein are selected from methanol, ethanol, isopropyl alcohol, n-butanol or n-propanol. The reaction is performed at 25°C to 500C, more preferably at 25°C to 30°C.
In yet another embodiment, the present invention provides a process for the purification of Bimatoprost comprising the steps of; a) dissolving Bimatoprost in a solvent, b) adding of an antisolvent, and c) isolating Bimatoprost.
According to the present invention, the solvent is selected from chlorinated solvents like dichloromethane, chloroform, 1,2-dichloroethane, carbon tetrachloride, or mixtures thereof.
The resulting solution is passed through a micron filter, like hyflow bed or silica gel bed. The anti-solvent is selected from diisopropyl ether and diethyl ether, preferably diisopropyl ether. The product may be isolated by the conventional techniques of filtration or centrifugation.
In yet another embodiment, the present invention provides a process for the preparation of triphenyl-4(phenyl-2-oxobutyl) phosphonium bromide (SC-1) as shown below

In yet another embodiment, the present invention provides a process for the preparation of triphenyl-4(phenyl-2-oxobutyl) phosphonium bromide (SC-1), comprising the steps of; a) treating benzyl acetone with bromine to obtain the bromo derivative. b) treating the bromo derivative with triphenyl phosphine in a solvent to get triphenyl- 4(phenyl-2- oxo butyl)phosphonium bromide.
According to the present invention benzyl acetone is reacted with alcoholic bromine solution in the presence of a solvent selected from methanol, ethanol, isopropanol, n-propanol or mixtures thereof.
The reaction of the bromo derivative with triphenyl phosphine to obtain triphenyl- 4(phenyl-2- oxobutyl)phosphonium bromide is performed in a solvent selected from toluene, tetrahydrofuran, diethyl ether or mixtures thereof. The major advantages of the present invention are summarized below: i. Bromobenzyl acetone is lachrymatory and hence, has not been isolated, ii. Addition of triphenyl-4(phenyl-2-oxobutyl)phosphonium bromide without isolating bromo benzyl acetone thus leading to very high yield, iii. Protection of both the hydroxy groups with tert-Butyldimethylsilyl chloride gives quantitative yields which are equivalent to theoretical yields, iv. Completion of the reaction in less than 2 hours using tert-Butyldimethylsilyl chloride without the formation of impurities, v. DIBAL reaction is either incomplete or forms more impurities when both the hydroxyl groups are unprotected or even one hydroxyl group is protected. When both the hydroxyl groups were protected with tert-Butyl Dimethyl silyl chloride, the
DIBAL reaction proceeds smoothly (with no formation of impurities) within 60-90 minutes with better yield and quality of product, vi. The pure product obtained after DIBAL reaction leads to the comparatively pure products obtained in the subsequent steps, vii. Use of aqueous ethyl amine solution, leads to incomplete reaction even after 80 hours whereas reaction in non-aqueous medium (by passing ethyl amine gas) leads to clean reaction and completion within 65-70 hours.
The invention is further illustrated by the following non-limiting examples.
EXAMPLES
Preparation of common intermediate: (1S, 5R, 6R, 7R)-6-[ (3S)-3-hydroxyl-5-phenyl-1E- pentenyl)-7-[ (4-phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one Step-1 : Preparation of Triphenyl-4(phenyl-2-oxo butyl) phosphonium bromide A) In a one litre round bottom flask fitted with an overhead stirrer, thermo pocket and dropping funnel was charged methanol (270ml) and benzyl acetone (100g) at 20 to 300C. Bromine solution (118.8 g bromine dissolved in 270 ml methanol) was added during 60 to 90 minutes maintaining internal temperature at 7 to10°C. The reaction mass was maintained at same temperature for 2 to 4 hours and checked HPLC for completion of reaction. D. M. water (674 ml) was added at 7 to 100C. The temperature was raised to 20 to 30° C and maintained for 12 hours at the same temperature. The organic layer was separated. The upper aqueous layer was extracted with dichloromethane (270ml) and combined with the organic layer. Dichloromethane was distilled out completely under reduced pressure to obtain viscous residue (140-150 g) which was dissolved in 300 ml toluene. B) Toluene (300ml) and triphenyl phosphine (167.5) was charged at 20 to 300C into a 2 litre round bottom flask equipped with an overhead stirrer, thermo pocket and dropping funnel. The toluene solution as obtained in step 1A was added at 20 to 300C. The reaction mass was maintained at the same temperature for about 2 to 4 hours and checked HPLC for completion of reaction. The solid was filtered and washed with toluene (150ml). The wet material obtained was taken in a flask and acetone (500ml) was charged into it and stirred for 2 hours at 20 to 300C. The product was filtered and washed with acetone (100ml). The wet material was dried under vacuum at 40 to 450C for 5 to 8 hours to obtain the title compound (200-220 g).
Step-2: Preparation of (1S,5R,6R,7R)-6-Oxo-5-phenyl-1E-pentenyl)-7-
[(phenylbenzoyl)oxy]-2-oxabicyclo[3.3.0] octane-3-one. Part- A): Preparation of Side chain Carbanion
2.9 g of potassium hydroxide was dissolved in 40 ml water at 20 to 300C into which triphenyl- 4(phenyl-2-oxobutyl)phosphonium bromide (16.5g) was charged along with toluene 77.5 ml) and dichloromethane (33ml) at 20 to 300C. The reaction mass was maintained under stirring for 3 hours at 20 to 300C. Sodium chloride (1.6 g) was added and further stirred for 15 minutes. Organic layer was separated and washed with brine solution. The solvent was distilled out completely under vacuum and further residue was dissolved in dichloromethane (30 ml) to get clear solution.
Part-B): Preparation of Corey aldehyde from Corey lactone.
Dimethoxy ethane (60ml) was charged into Corey lactone (10 g) at 25 to 30°c in a 250 ml four neck round bottom flask equipped with mechanical stirrer and dropping funnel. The suspension was stirred for 10 minutes, charged dicyclohexyl carbodiimide (23.3) and dimethyl sulfoxide (DMSO) (17.75 g) at 25 to 300C. The reaction mass was cooled to 20 to 220C and phosphoric acid was added (1.65g). The reaction mixture was maintained at 20 to 22° C for about 20 to 30 minutes. The temperature was raised to 25 to 300C. After completion of reaction, the unwanted dicyclohexyl urea was filtered and washed with dimethoxy ethane (20 ml). The filtrate contains Corey aldehyde.
Part C: Preparation of (1S,5R,6R,7R )-6-oxo-5-phenyl-1 E-Penten) -7-[ (phenyl benzoyl) oxy]-2-oxabicyclo [3.3.0] octane -3-one.
The filtrate (Corey aldehyde) was taken in a separate flask and slowly the reaction mass was added from Step 2(B) at 25 to 30°c. The reaction mass was maintained at 25 to 30°C for one hour. Raised the temperature to 35 to 400C and stirred for 2 to 3 hours till reaction completes.
The reaction mass was cooled to 18 to 2O0C and dilute HCI (2.5 ml cone. HCL diluted in 17.5 ml water) was added to it. The reaction mass was stirred for 15 to 20 minutes, filtered, and the organic layer was separated. The organic layer was washed twice with water (2X50 ml). The solvent was distilled out completely under vacuum and the residue was charged with ethanol
(50 ml). The temperature was raised to 40 to 450C and stirred for 15 minutes to get clear solution. The solution was gradually cooled to 20°C and further to 0 to 5°C. The stirring was continued for 2 hours at 0 to 5°Cfollowed by filtration and washing with 20 ml cold ethanol. 7-8 g of the title compound was obtained. (HPLC purity: > 98%)
Step-3: Preparation of (1S, 5R, 6R, 7R)-6-[ (3S)-3- hydroxyl-5-phenyl-1E-pentenyl)-7-[ (4- phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one.
Charged (-) DIP chloride ( 40 gm 50 % solution ) to 250 ml round bottom flask fitted with overhead stirrer, thermo pocket and a dropping funnel at 25 to 300C . The reaction mass was cooled to -20 to -25°C. 10 g (1S,5R,6R,7R)-6-oxo-5-phenyl-1E-pentenyl)-7- [(phenylbenzoyOoxyJ^-oxabicycloβ.S.OJoctane-S-one in 50 ml tetrahydrofuran was slowly added for 45 to 60 minutes at -20 to 25°C. Stirring was continued at -20 to -25° C for 5 to 8 hours. After completion of reaction, methanol (40 ml) was added at -20 to -25°C. The temperature was raised to - 20 to 30°C and stirred for 60 minutes. The solvents were distilled out completely under vacuum at 40 to 45°C. Dichloromethane (50ml) was added at 25°C and stirred to get clear solution. The solution was washed with ammonium chloride solution (3 x 60ml) and finally with brine solution (50 ml), heated to reflux (650C) for 2 to 3 hours. The reaction mass was cooled to 25 to 300C and maintained for 8 to 10 hours.. It was further cooled to 0 to 5°C and stirred for 2 hours at 0 to 5°C. The solid was filtered and washed with 10 ml of cold methanol to get 5-6 g of the title compound. (HPLC purity: 99%, Chiral purity: 97- 99.5%)
Step-3A: Preparation of ("IS, 5R, 6R, 7R)-6-[ (3S)-3- hydroxyl-5-phenyl-1E-pentenyl)-7-[ (4-phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one. Charged (-) DIP chloride ( 55.4 gm 50 % solution ) to 250 ml round bottom flask fitted with overhead stirrer, thermo pocket and a dropping funnel at 25-30°C . Cooled the reaction mass - 30 to -35°C Slowly added 10 g 1S,5R,6R,7R )-6-oxo-5-phenyl-1E-pentenyl)-7-[ (phenyl benzoyl)oxy]-2-oxabicyclo [3.3.0] octane -3-one in 50 ml THF for 45-60 minutes at -30 to - 35°C. Maintained under stirring at -30 to -35° C for 5- 8 hours. After completion of reaction, added acetone (35 ml) maintaining temperature -30 to 35°C. Raised temperature to 0-50C and stirred 10-12 hours at 0-50C. Adjust pH to 5-7 with sodium hydroxide solution. Separate the layer and aqueous layer was extracted with dichloromethane. Combined organic layer was washed with water fallowed by brine solution. Distilled out solvents completely u/v at 35°C. The reaction mass was cooled to 20-250C and charged n-hexane (100 ml). Maintain for 12 hours at 20-250C. Further cooled reaction mass to 0-5°C and maintained for 2 hours. Filtered crude product was stirred in MDC (80 ml) and undissolved material (salts) was filtered. Taken filtrate and distilled out MDC u/v. To the viscous residue charged n-hexane and maintained for 2-3 hours at 20-25°C. Further cooled to 0-5°C and maintain for 2 hours at 0-5°C. Filter the product and washed with n-hexane. Dried at 45°C to get 7-9 g title compound. Note: The product was crystallized in a mixture of methanol and diisopropyl ether.
Example 1 : Preparation of Latanoprost
Step 1: Preparation of 1S,5R,6R,7R)-6-[(3R)-3-Hydroxy-5-phenyl-1pentyl)-7-(4- phenyl benzoyl)oxy}-2-oxabicyclo[3.3.0] octan-3-one
1N sodium hydroxide (12.5 ml), ethanol (300 ml), 5% Palladium on carbon (5g) and the product obtained in step 3 (25 g) were charged into autoclave vessel at 20 to 300C. The reaction mixture was stirred at 25 to 3O0C under hydrogen pressure (4-5 kg) till reaction is complete (5 to 8 hours). After completion of reaction, the unreacted catalyst was filtered off and washed with ethanol. Ethanol was distilled out completely under vacuum at 40 to 45°C. Ethyl acetate (250 ml) and 1 N hydrochloric acid (12.5 ml) were added to the residue at 25 to 3O0C, stirred for 20 minutes followed by the separation of ethyl acetate layer. The aqueous phase was extracted twice with ethyl acetate. The combined ethyl acetate layers were washed with brine solution (100 ml). Ethyl acetate was distilled out completely under vacuum to obtain viscous residue (-24 g) which was proceeded to next step without purification (HPLC purity >97%) Step 2: Preparation of 1S,5R,6R,7R)-6-[(3R)-3-Hydroxy-5-phenyl-1-pentyl)-7-(S)- hydroxy-2-oxabicyclo[3.3.0] octan-3-one
To a solution of compound obtained in step 1 of example-1 (25 g) in methanol (250 ml) was added potassium carbonate (4.65 g) at 20 to 300C. The reaction mixture was stirring at 20 to 30°C for 5 to 7 hours. After the completion of the reaction, methanol was distilled out completely under vacuum at 40 to 45°C. 1 N hydrochloric acid (320 ml) was charged along with ethyl acetate (250 ml) and stirred for 30 minutes at 20 to 30°C. The organic layer was separated and the aqueous layer was extracted with ethyl acetate (250 ml). The combined organic layer was distilled out under vacuum at 40 to 45°C to remove ethyl acetate completely. The residue thus obtained was separated by column chromatography to obtained 13.5 to 15 g title compound as viscous oil (HPLC purity: 99%)
Step 3: Preparation of 1S,5R,6R,7R)-6-[(3S)dimethyltert-butylsilyloxy-5-phenyl-1- pentyl)(7S)dimethyltert-butylsilyloxy-2-oxabicyclo[3.3.0]octan-3-one
To a 500 ml round bottom flask fitted with a mechanical stirrer and thermo pocket, was charged tetrahydrofuran (170 ml) and (1S,5R,6R,7R)-6-[(3R)-3-Hydroxy-5-phenyl-1pentyl)-7- (S)-hydroxy-2-oxabicyclo[3.3.0]octan-3-one (17 g) was added at 25 to 30°C. The reaction mixture was stirred for 10 minutes and imidazole (22.9 g) was added to it followed by tert- Butyldimethylsilyl chloride (33.6 g) at 25 to 300C. The reaction mixture was heated at 65 to 700C for 60 to 90 minutes for completion of reaction. The reaction mass was cooled to 40 to 45°C and tetrahydrofuran was distilled out completely under vacuum. Dichloromethane (170 ml) was charged at 25 to 300C and stirred for 10 minutes to get the clear solution. The
reaction mass was washed with 10 % citric acid (50 ml). The reaction mass was filtered through filter cloth and solvent was distilled out completely under vacuum at 40 to 450C. Methyl tert-Butylether (MTBE) (50 ml) was charged to the residue and stirred for 15 minutes to get clear solution. MTBE was distilled out completely under vacuum at 40 to 45°C to get 27-29 g of title compound as a white semi solid (HPLC purity 96 - 97%)
Step 4: Preparation of 1S,5R,6R,7R)-6-[(3R)dimethyl tert-butylsilyloxy-5-phenyl-1-pentyl) (7S) dimethyl t.butyl silyloxy -2-oxabicyclo[3.3.0] octan-3-ol
Toluene (320 ml) was charged into the compound isolated in step 3 of example-1 (32 g) in a 500 ml round bottom flask equipped with overhead stirrer, thermo pocket and dropping funnel at 25 to 30°. The solution was cooled to -700C and slowly added Diisobutyl aluminum hydride (1 M solution in toluene, 130 ml) maintaining the temperature at -70 to -800C. The reaction mixture was stirred at same temperature for 90 min and progress of reaction was monitored by TLC. After the completion of the reaction, methanol (130 ml) was charged at -70 to -80°C. Raised the temperature from 25 to 300C and maintained for 30 minutes. The unwanted salt was filtered and washed with methanol (30 ml). The filtrate was passed through silica column and solvent was distilled out completely under vacuum to obtained above said product as viscous oil (33 g) (HPLC purity - 93%)
Step 5: Preparation of (Z)-7-(1R,2R,3R,5S)-5-hydroxy-2-[(3R)dimethyltert-butyl silyloxy- 5-(phenyl-1 -pentyl)-3-(dimethyl tert-butylsilyloxy)cyclopentyl-5-hetp-enoic acid.
4-Carboxybutyl)triphenylphosphonium bromide (66.2 g) was suspended in tetrahydrofuran (265 ml) at 25 to 30°C. The reaction mass was cooled to 15 to 20°C under stirring and lot wise potassium tert-butoxide (33.5 g) was added at 15 to 200C over a period of 30 minutes. The temperature was raised to 20 to 25° C, maintained at same temperature for 30 minutes. The reaction mass was cooled to -10 to -15°C and solution of compound of step 4 of example 1 (10g in 30 ml tetrahydrofuran) was added slowly at -10 to -15°C. The reaction mixture was maintained at -10 to -15°C for 2 hours until TLC shows the reaction was complete. Tetrahydrofuran was distilled out completely under vacuum below 300C. Water (100 ml) was charged and stirred for 30 minutes at 25 to 3O0C. The unwanted salt was filtered and washed with water (10 ml). To the filtrate (10%) citric acid solution was added to adjust pH to 4.0. Ethyl acetate (250 ml) was added and stirred for 20 minutes. The organic layer was separated and the aqueous layer was extracted with ethyl acetate (250 ml). The combined ethyl acetate layers were washed with brine solution and distilled out completely under vacuum at 40-45°C.
The crude product thus obtained was purified by column chromatography to get 7-8 grams title compound.
Step 6: Preparation of lsopropyl (Z)-7-(1RI2R,3R,5S)-5-hydroxy-2-[(3R)dimethyl tert- butylsilyloxy-5-(phenyl-1-pentyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5-hetpanoate Dimethyl formamide (100 ml) and compound of step 5 of example 1 (10 g) were taken in a 250 ml round bottom flask at 25 to 30°C. Cesium carbonate (7.8 g) was charged under stirring at 25 to 30°C and stirred for 15 minutes. Further isopropyl bromide (3.9 g) was added and temperature was raised to 40 to 45°C. The temperature was maintained at 40 to 45°C for 3 to 4 hours until reaction is completed by TLC. The reaction mass was cooled to 25 to 300C and water (100 ml) was charged. The pH of the solution was adjusted with potassium hydrogen sulphate (10% solution, ~20 ml). Charged ethyl acetate (100 ml) and stirred for 15 to 20 minutes. Separated the organic layer and aqueous layer was extracted with ethyl acetate. The combined organic layers were washed with brine solution. Finally organic layer was passed through silica column and distilled out solvent completely under vacuum at 40 to 45°C to get viscous residue (10 g) as a protected Latanoprost.
Step 7: Preparation of Latanoprost
In a 250 ml round bottom flask fitted with mechanical stirrer, thermo pocket and stopper, charged isopropyl alcohol (30 ml) and compound obtained in step 6 of example 1 (10 g) at 25 to 30°C. 2N HCI (40 ml) was slowly added during 25 to 30 minutes at 25 to 300C. The mixture was stirred at 25 to 300C for 3 to 5 hours till the reaction is completed by TLC. Water (100 ml) was charged and pH adjusted to 6.5 with sodium bicarbonate solution at 25 to 3O0C. Charged ethyl acetate (150 ml) and stirred for 20 minutes. The organic layer was seperated and aqueous layer was extracted twice with ethyl acetate. The ethyl acetate fractions were combined and washed with brine solution. The solvent was distilled out completely under vacuum below 300C. The crude product was purified by column chromatography/ HPLC to obtain 5 g Latanoprost.
Example 2: Preparation of Bimatoprost Step 1 : Preparation of 1(S) -2oxa-3-oxo-6R-(3S-hydroxy-5- phenyl-1- trans-pentenyl)-7- R-hydroxy-cis-bicyclo- [3.3.0]-octane
To a solution of product obtained in step 3 (10 g) in methanol (100 ml) was charged potassium carbonate (1.86 g) at 20-30°C and stirred at 20 to 30°C for 5 to 7 hours. After the completion of the reaction, methanol was distilled out completely under vacuum at 40 to 45°C. 1N Hydrochloric acid (120 ml) and ethyl acetate (100 ml) were charged and stirred for 30 minutes
at 20 to 30°C. The organic layer was separated and the aqueous layer was extracted with ethyl acetate twice. The combined ethyl acetate layers were washed with brine solution and ethyl acetate was distilled out completely under vacuum at 40 to 45°C. The residue thus obtained was separated by column chromatography to obtain 5.4 to 6.0 g title compound as viscous oil.
Step 2: Preparation of 1S,5R,6R,7R)-6-[(3S)dimethyltert-butylsilyloxy-5-phenyl-1- pentenyl) (7S)dimethyltert-butyl silyloxy-2-oxabicyclo[3.3.0]octan-3-one To a 500 ml round bottom flask fitted with mechanical stirrer and thermo pocket, charged tetrahydrofuran (170 ml) and 1S,5R,6R,7R-6-[(3R)-3-hydroxy-5-phenyl-1-pentenyl)-7-(S)- hydroxy-2-oxabicyclo [3.3.0] octan-3-one (16 g) at 25 to 30°C. The reaction mixture was stirred for 10 minutes followed by the addition of imidazole (21.6 g) and tert-Butyld imethylsilyl chloride (33.6 g) at 25 to 30°C. The temperature was raised to 65 to 700C and maintained for 60 to 90 minutes until the completion of the reaction. The reaction mass was cooled to 40 to 45°C and tetrahydrofuran was distilled out completely under vacuum. Dichloromethane (160 ml) was charged at 25 to 300C and stirred for 10 minutes to get the clear solution. The reaction mass was washed with 10% citric acid (50 ml). The solvent was distilled out completely under vacuum at 40 to 45°C. Methyl tert-Butyl ether (MTBE) (50 ml) was charged to the residue and stirred for 15 minutes to get clear solution. MTBE was distilled out completely under vacuum at 40 to 45°C to get 27-29 g of title compound as a white residue.
Step 3: Preparation of 1S,5R,6R,7R)-6-[(3R)dimethyltert-butylsilyloxy-5-phenyl-1- pentenyl) (7S) dimethyltert-butyl silyloxy-2-oxabicyclo[3.3.0] octan-3-ol Toluene (260 ml) was charged into the compound isolated in step 2 of example 2 (25 g) in a 500 ml round bottom flask equipped with overhead stirrer, thermo pocket and dropping funnel at 25 to 3O0C. The reaction mixture was cooled to -700C and slowly added Diisobutyl aluminum hydride (122 ml) (20% solution in toluene) maintaining the temperature at -70 to - 80°C. Stirred at same temperature for 45 minutes and the progress of reaction was monitored was by TLC. Methanol (105 ml) was charged at -70 to -80°C after completion of the reaction. The temperature was raised to 25 to 3O0C and maintained for 30 minutes. The unwanted salt was filtered and and washed with methanol (50 ml). The filtrate was passed through silica column and solvent was distilled out completely under vacuum to obtain the above said product as viscous oil (23 g).
Step 4: Preparation of (Z)- 7-(1R,2R,3R,5S) -5-hydroxy -2- [(3R) dimethyl tbutyl silyloxy - 5-(phenyl-1-pentenyl)-3-( dimethyl t.butyl silyloxy) cyclopentyl -5-hetp-enoic acid.
(4-Carboxybutyl)triphenylphosphonium bromide (151.5 g) was suspended in tetrahydrofuran (604 ml) at 25 to 300C. The reaction mixture was cooled under stirring to 15 to 200C and lot wise potassium tertiary butoxide (76.8 g) was added at 15 to 20° C and maintained for 30 minutes. Cooled slowly to -10 to -15° C and a solution of compound obtained in step 3 of example 2 (22.8 g in 69 ml tetrahydrofuran) was added slowly at -10 to -15°C. Maintained at - 10 to -15°C for 2 hours until TLC shows the reaction was complete. The tetrahydrofuran was distilled out completely under vacuum below 30°C. Water (228 ml) was charged and stirred for 30 minutes at 25 to 300C. The unwanted salt was filtered and washed with water (25 ml). To the filtrate added (10%) citric acid solution to adjust the pH to 4.0. Ethyl acetate (450ml) was added and stirred for 20 minutes. The organic layer was seperated. The aqueous layer was extracted with ethyl acetate (2x150 ml). Combined all the ethyl acetate layers and washed with brine solution. Ethyl acetate was distilled out completely under vacuum at 40 to 45°C. The crude product thus obtained was purified by column chromatography to get 18- 2O g title compound. Step 5: Preparation of Methyl (Z)-7-(1R,2R,3R,5S)-5-hydroxy-2-[(3R)dimethyl tert- butylsilyloxy-5-(phenyl-1-pentenyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5- hetpanoate
Dimethyl formamide (100 ml) and compound isolated in step 4 of example 2 (10 g) was taken in 250 ml round bottom flask at 25 to 30°C. Cooled to 0 to 10°C. Cesium carbonate (7.9 g) was added under stirring at 0 to 100C and stirred for 15 minutes followed by the addition of methyl iodide (3.4 g) at 0 to 10°C. The temperature was maintained at 0 to 10°C for 3 to 4 hours until the reaction is completed by TLC. Water (400 ml) was added and the pH was adjusted 6.5 to 7.0 using potassium hydrogen sulphate (10% solution, 20 ml). Ethyl acetate (100 ml) was charged and stirred for 15 to 20 minutes. The organic layer was separated and aqueous layer was extracted with ethyl acetate (2x75 ml). The combined organic layers were washed with brine solution. Finally, organic layer was washed with brine solutionand the solvent was distilled out completely under vacuum at 40 to 45°C to get viscous residue (9 g).
Step 6: Preparation of (Z)-7-(1R,2R,3RI5S)-5-hydroxy-2-[(3R)dimethyltert-butyl silyloxy- 5-(phenyl-1-pentenyl)-3-(dimethyltert-butylsilyloxy)cyclopentyl-5-N-ethyl heptenamide (protected Bimatoprost)
Methanol (25 ml) and compound isolated in step 5 of example 2 (2.3 g) were taken in 100 ml round bottom flask at 25 to 300C. The reaction mass was cooled to 10 to 15°C, Purged ethyl amine gas for 3 to 4 hours. The temperature of the reaction mixture was increased to 25 to 30°C and maintained at 25 to 30°C for 65 to 70 hours until reaction is completed by TLC. The
solution was concentrated at 30 to 35°C, and D.M water (25 ml) was charged to the residue. The pH was adjusted to 6.5 to 7.0 with potassium hydrogen sulphate solution. Ethyl acetate (25 ml) was added and stirred for 15 minutes. The organic layer was separated. The aqueous layer was extracted with ethyl acetate (2x20 ml). The combined organic layer was washed with brine solution (30ml). The solvent was distilled out completely under vacuum at 35 to 40°C to get clear viscous residue (2-2.3 g) as a protected Bimatoprost.
Step 7: Preparation of Bimatoprost
In a 100 ml round bottom flask fitted with stirrer, thermo pocket and stopper was charged methanol (20 ml) and the compound obtained in step 6 of example 2 at 25 to 300C. The resulting mixture was cooled to 15 to 200C and slowly added 2N HCI (5 ml) at 15 to 200C. The reaction mixture was maintained under stirring at 15 to 200C for 3 to 5 hours till reaction is completed by TLC. Water (40 ml) was added and cooled to 0 to 100C, the pH was adjusted to 6.5 to 7.0 with 5% sodium carbonate solution at 0 to 100C. Ethyl acetate (40 ml) was charged and stirred for 15 minutes. The organic layer was separated and aqueous layer extracted with ethyl acetate (2x 40 ml). The combined ethyl acetate fraction was washed with brine solution. The solvent was distilled out completely under vacuum below 30°C. The crude product was purified by flash column chromatography/ preparative HPLC to obtained pure Bimatoprost. (0.8-1.O g) Step 8: Purification of Bimatoprost
The crude Bimatoprost (1.2 g) was dissolved in dichloromethane (15 ml) at 20-25°C to get clear solution. The reaction mass was filtered through micron filter to make particle free. Charged filtrate to RB flask and slowly added diisopropyl ether (90 ml) over a period of 30-45 minutes. The product was filtered and washed with diisopropyl ether to obtain 0.8-0.9 g pure Bimatoprost.
Step 8A: Purification of Bimatoprost
The crude product obtained in step 7 (1.2 g) was dissolved in dichloromethane (15 ml) to get clear solution at 25 to 300 C. slowly added diisopropyl ether (90 ml) over a periodof 30 to 45 minutes. Stirred for one hour at 25 to 300 C, filtered and washed with diisopropyl ether to obtain 0.8 to 0.9 g pure bimatoprost
Note: In the purification step, diethyl ether can also be used instead of diisopropyl ether.
Claims
1. A process for the synthesis of a compound of formula 1 :

Formula 1 comprising the steps of; a) oxidizing the compound of formula 2,

Formula 2
Wherein, R is PPB, benzoyl, Cbz or BOC. to form a compound of formula 3,

Formula 3 b) reacting a compound of formula 3, with triphenyl-4-(phenyl-2- oxobutyl) phosphonium bromide having the formula SC-1,

SC-1 in presence of a solvent to form a compound of formula 4,

Formula 4 c) reducing the compound of formula 4 in presence of (-)-Diisopinocampheyl chloroborane to form a compound of formula 5,

Formula 5 d) optionally, hydrogenating the double bond of the compound of formula 5 in the presence of catalyst in a solvent to form a compound of formula 6,

Formula 6 e) deprotecting a compound of formula 6 in a base, solvent to form a compound of formula 7,

Formula 7 f) protecting the hydroxyl groups of the compound of formula 7 with a tert- butyldimethylsilyl chloride, in the presence of a base to form a compound of formula 8

Formula 8 g) reducing the lactone group in the compound of formula 8 to obtain a di-protected lactol of formula 9, and
TBD 
Formula 9 h) reacting the compound of formula 9 with 4-carboxy butyltriphenyl phosphonium bromide having the formula :  in the presence of a base and a solvent to generate the compound of formula 1.
2. The process according to claim 1(a) wherein, the oxidation is carried out in a combination of dimethylsulfoxide and dicyclohexylcarbodiimide.
3. The process according to 1(b) wherein, the solvent is selected from dimethoxyethane or toluene.
4. The process according to claim 1(d) wherein, the hydrogenation of compound of formula 5 is performed using a catalyst selected from palladium on carbon, platinum or nickel,
5. The process according to claim 1(d) wherein, the solvent is selected from ethanol, methanol, isopropanol or n-butanol.
6. The process according to claim 1(e) wherein, the deprotection of the compound of formula 6 is performed using a base selected from potassium carbonate, sodium carbonate or cesium carbonate.
7. The process according to claim 1(e) wherein, the solvent is selected from ethanol, methanol, isopropanol or n-butanol.
8. The process according to claim 1(f) wherein, the base is selected from imidazole, triethylamine, diisopropylethylamine or tributylamine.
9. The process according to claim 1(g) wherein, the reduction of the lactone group is carried out using diisobutyl aluminum hydride.
10. The process according to claim 1(h) wherein, the base is selected from n-butyl lithium, sodium amide, sodium hydride, sodium methoxide, sodium ethoxide, potassium tert- butoxide or potassium ethoxide, in the presence of a solvent selected from tetrahydrofuran, toluene or dichloromethane.
11. A process for the synthesis of Latanoprost,
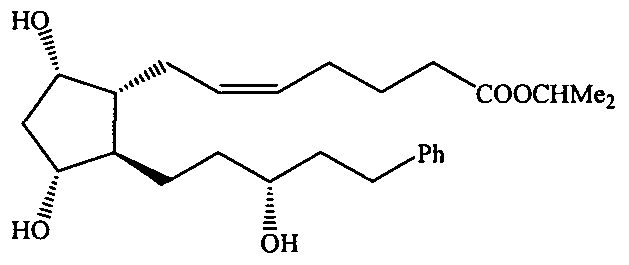

Formula 1 wherein the dotted bond ( ) is no bond, with isopropyl bromide in the presence of a base to form a compound of formula 10, and
TB 
■ Formula 10 b) deprotecting the silyl protected diol to give Latanoprost.
12. The process according to claim 10(a) wherein, the base is selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate or sodium bicarbonate.
13. The process according to claim 10(b) wherein, the deprotection of the silyl protecting group is carried out in the presence of an acid selected from an aqueous hydrochloric acid or aqueous sulfuric acid.
14. A process for the synthesis of Bimatoprost,


Formula 1 wherein dotted bond ( ) is a single bond, with methyl iodide in the presence of a base to form a compound of formula 11 ,

Formula 11 b) reacting the ester derivative of formula 11 with ethylamine in a solvent, and

Formula 12 c) deprotecting the silyl protected diol to give Bimatoprost.
15. The process according to claim 14(a) wherein, the base is selected from cesium carbonate, potassium carbonate, sodium carbonate, potassium bicarbonate or sodium bicarbonate.
16. The process according to claim 14(b) wherein, the amidification is carried out in a solvent selected from methanol, ethanol, isopropanol, n-propanol, n-butanol, isobutanol or mixtures thereof.
17. The process according to claim 14(c) wherein, the deprotection of the silyl group is carried out in the presence of an acid.
18. The process according to claim 17 wherein, the acid is selected from aq. hydrochloric acid or aq. sulfuric acid.
19. A process for the purification of Bimatoprost comprising the steps of; a) dissolving Bimatoprost in a solvent, b) adding an antisolvent, and c) isolating Bimatoprost.
20. The process according to claim 19 wherein, the solvent is selected from dichloromethane, chloroform, 1 ,2-dichloroethane, carbon tetrachloride,
21. The process according to claim 19 wherein, the anti-solvent is selected from diisopropyl ether or diethyl ether.
22. A process for the synthesis of triphenyl-4-(phenyl-2-oxobutyl)phosphonium bromide, used in claim 1(b) of formula SC-1,

23. The process according to claim 21 (a) wherein, benzyl acetone is reacted with alcoholic bromine in a solvent selected from methanol, ethanol, isopropanol, n-propanol, preferably, methanol.
24. The process according to claim 21 (b) wherein, the solvent is selected from toluene, tetrahydrofuran, diethyl ether, preferably toluene.
25. The process as in all the preceeding claims wherein Latanoprost and Bimatoprost are having purity greater than 99.8%.
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| IN121/CHE/2009 | 2009-01-19 | ||
| IN121CH2009 | 2009-01-19 |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| WO2010109476A2 true WO2010109476A2 (en) | 2010-09-30 |
| WO2010109476A3 WO2010109476A3 (en) | 2012-10-04 |
Family
ID=42781614
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/IN2010/000026 Ceased WO2010109476A2 (en) | 2009-01-19 | 2010-01-18 | Improved process for the preparation of prostaglandins and analogues thereof |
Country Status (1)
| Country | Link |
|---|---|
| WO (1) | WO2010109476A2 (en) |
Cited By (8)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2012146085A1 (en) * | 2011-04-29 | 2012-11-01 | 上海源力生物技术有限公司 | Intermediate for synthesizing prostaglandin medicines and preparation method therefor |
| CN103604895A (en) * | 2013-12-05 | 2014-02-26 | 武汉武药科技有限公司 | Method for analytical separation of (-) benzoyl Corey lactone optical isomers by HPLC |
| US8901319B2 (en) | 2009-10-16 | 2014-12-02 | Cayman Chemical Company, Incorporated | Process for the preparation of F-series prostaglandins |
| JP2015083570A (en) * | 2011-08-24 | 2015-04-30 | チャイロゲート インターナショナル インク.Chirogate International Inc. | Intermediate for synthesis of benzoindene prostaglandin and manufacturing method therefor |
| WO2015136317A1 (en) | 2014-03-13 | 2015-09-17 | CHINOIN Gyógyszer és Vegyészeti Termékek Gyára Zrt. | New process for the preparation of high purity prostaglandins |
| EP3235810A1 (en) | 2016-04-19 | 2017-10-25 | Gentec, S.A. | Process for the preparation of bimatoprost |
| CN112481313A (en) * | 2020-11-23 | 2021-03-12 | 江苏阿尔法药业有限公司 | Enzymatic synthesis method of bemepiride intermediate |
| CN112608294A (en) * | 2020-12-16 | 2021-04-06 | 西安国康瑞金制药有限公司 | Preparation method of latanoprost |
Family Cites Families (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2008037782A (en) * | 2006-08-04 | 2008-02-21 | Daiichi Fine Chemical Co Ltd | Method for producing prostaglandine derivative |
-
2010
- 2010-01-18 WO PCT/IN2010/000026 patent/WO2010109476A2/en not_active Ceased
Cited By (13)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US8901319B2 (en) | 2009-10-16 | 2014-12-02 | Cayman Chemical Company, Incorporated | Process for the preparation of F-series prostaglandins |
| WO2012146085A1 (en) * | 2011-04-29 | 2012-11-01 | 上海源力生物技术有限公司 | Intermediate for synthesizing prostaglandin medicines and preparation method therefor |
| JP2015083570A (en) * | 2011-08-24 | 2015-04-30 | チャイロゲート インターナショナル インク.Chirogate International Inc. | Intermediate for synthesis of benzoindene prostaglandin and manufacturing method therefor |
| CN103604895A (en) * | 2013-12-05 | 2014-02-26 | 武汉武药科技有限公司 | Method for analytical separation of (-) benzoyl Corey lactone optical isomers by HPLC |
| CN103604895B (en) * | 2013-12-05 | 2015-04-22 | 武汉武药科技有限公司 | Method for analytical separation of (-) benzoyl Corey lactone optical isomers by HPLC |
| US10501410B2 (en) | 2014-03-13 | 2019-12-10 | CHINOIN Gyógyszer és Vegyészeti Termékek Gyára Zrt. | Pocess for the preparation of high purity prostaglandins |
| WO2015136317A1 (en) | 2014-03-13 | 2015-09-17 | CHINOIN Gyógyszer és Vegyészeti Termékek Gyára Zrt. | New process for the preparation of high purity prostaglandins |
| EP3235810A1 (en) | 2016-04-19 | 2017-10-25 | Gentec, S.A. | Process for the preparation of bimatoprost |
| WO2017182465A1 (en) | 2016-04-19 | 2017-10-26 | Gentec, S.A. | Process for the preparation of bimatoprost |
| US10626080B2 (en) | 2016-04-19 | 2020-04-21 | Gentec, S.A. | Process for the preparation of bimatoprost |
| CN112481313A (en) * | 2020-11-23 | 2021-03-12 | 江苏阿尔法药业有限公司 | Enzymatic synthesis method of bemepiride intermediate |
| CN112608294A (en) * | 2020-12-16 | 2021-04-06 | 西安国康瑞金制药有限公司 | Preparation method of latanoprost |
| CN112608294B (en) * | 2020-12-16 | 2021-10-26 | 西安国康瑞金制药有限公司 | Preparation method of latanoprost |
Also Published As
| Publication number | Publication date |
|---|---|
| WO2010109476A3 (en) | 2012-10-04 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| US7498458B2 (en) | Process for the preparation of prostaglandins and analogues thereof | |
| US20040171873A1 (en) | Process for the preparation of 17-phenyl-18,19,20-thinor-pgf 2a and its derivatives | |
| US8772544B2 (en) | Process for the production of bimatoprost | |
| KR101561171B1 (en) | Process for the preparation of f-series prostaglandins | |
| NO317060B1 (en) | Fluorine-containing prostaglandin derivatives, their use and medicaments containing the derivative | |
| US20150011755A1 (en) | Amine salts of prostaglandin analogs | |
| EP2496553A1 (en) | A novel process for the preparation of prostaglandins and intermediates thereof | |
| KR20140107541A (en) | Process for the preparation of travoprost | |
| WO2012011128A1 (en) | Preparation of prostaglandin derivatives | |
| KR101522218B1 (en) | Processes and intermediates for the preparations of prostaglandins | |
| CN108084073B (en) | A method of purifying Bimatoprost | |
| EP0471856B1 (en) | 15-deoxyprostaglandin derivative | |
| JP2008037782A (en) | Method for producing prostaglandine derivative | |
| US20080033176A1 (en) | Method for preparing prostaglandin derivative | |
| EP3445745B1 (en) | Process for the preparation of bimatoprost | |
| CN116947725A (en) | Synthesis method of travoprost | |
| KR20250050531A (en) | Method for producing Tafluprost | |
| AU2002309221A1 (en) | Preparation of 17-phenyl-18,19,20- trinor-PGF2a and its derivatives |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 10755537 Country of ref document: EP Kind code of ref document: A2 |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| 122 | Ep: pct application non-entry in european phase |
Ref document number: 10755537 Country of ref document: EP Kind code of ref document: A2 |