WO2004065360A2 - Piperazine derivatives and their use as synthesis intermediates - Google Patents
Piperazine derivatives and their use as synthesis intermediates Download PDFInfo
- Publication number
- WO2004065360A2 WO2004065360A2 PCT/EP2004/000399 EP2004000399W WO2004065360A2 WO 2004065360 A2 WO2004065360 A2 WO 2004065360A2 EP 2004000399 W EP2004000399 W EP 2004000399W WO 2004065360 A2 WO2004065360 A2 WO 2004065360A2
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- compound
- formula
- reaction
- compounds
- group
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 C*(C*(C)OCC(*)=O)N(CC1)CCN1C(c1ccccc1)c(cc1)ccc1Cl Chemical compound C*(C*(C)OCC(*)=O)N(CC1)CCN1C(c1ccccc1)c(cc1)ccc1Cl 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D271/00—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms
- C07D271/02—Heterocyclic compounds containing five-membered rings having two nitrogen atoms and one oxygen atom as the only ring hetero atoms not condensed with other rings
- C07D271/06—1,2,4-Oxadiazoles; Hydrogenated 1,2,4-oxadiazoles
- C07D271/07—1,2,4-Oxadiazoles; Hydrogenated 1,2,4-oxadiazoles with oxygen, sulfur or nitrogen atoms, directly attached to ring carbon atoms, the nitrogen atoms not forming part of a nitro radical
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D295/00—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms
- C07D295/04—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms
- C07D295/14—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
- C07D295/145—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals with the ring nitrogen atoms and the carbon atoms with three bonds to hetero atoms attached to the same carbon chain, which is not interrupted by carbocyclic rings
- C07D295/15—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals with the ring nitrogen atoms and the carbon atoms with three bonds to hetero atoms attached to the same carbon chain, which is not interrupted by carbocyclic rings to an acyclic saturated chain
Definitions
- the present invention relates to new piperazine derivatives and to their use as synthesis intermediates, especially for the preparation of pharmaceutically active compounds.
- Levorotatory [2-[4-[(4-chlorophenyl)phenylmethyl]-l-piperazinyl]ethoxy]acetic acid also known by the generic name of levocetirizine, has been proven useful as therapeutic agent for the treatment of allergic diseases.
- Said compound may be obtained from its racemic inixture by resolution of the enantiomers of cetirizine ([2-[4- [(4-chlorophenyl)phenylmethyl]-l-piperazinyl]ethoxy]acetic acid).
- GB 2,225,321 describes a process for the preparation of cetirizine in the levorotatory form, dextrorotatory form or a mixture thereof comprising the hydrolysis of enantiomerically pure or racemic [2-[4-[(4-chlorophenyl)phenylmethyl]-l- piperazinyl]ethoxy]-acetonitrile.
- the present invention relates to enantiomerically pure compounds of formula (I), in free form or in salt form,
- Y is hydroxy or a leaving group and n is 1, 2, 3, 4 or 5.
- Compounds of formula (I) include a centre of asymmetry indicated by the asterisk.
- enantiomerically pure compounds refers to compounds containing at least 90% of one enantiomer ((R) or (S)), preferably at least 98%, of the total amount of both enantiomers.
- leaving group has the same meaning to the skilled man (Advanced Organic Chemistry: reactions, mechanisms and structure - Third Edition by Jerry March, John Wiley and Sons Ed.; 1985 page 179) and represents a group which is part of and attached to a substrate molecule ; in a reaction where the substrate molecule undergoes a displacement reaction (with for example a nucleophile), the leaving group is then displaced.
- Preferred leaving groups are halogen, sulfonic esters such as OS ⁇ 2-C ⁇ H -CH3 (p-toluenesulfonate), OS ⁇ 2-C ⁇ H 4 -Br (p-bromobenzenesulfonate), OS ⁇ 2-CeH -N ⁇ 2 (p- nitrobenzenesulfonate), OSO2-CH3 (methanesulfonate), OSO2-CF3 (trifluoromethanesulfonate), OSO2-C1F9 (nonafluorobutanesulfonate), OSO2-CH 2 -CF3 (2,2, 2-trifluoroethanesulfonate) , OSO2- (CH2) n -NMe 3+ (axnmoniumalkanesulfonate) , OSO 2 -F (fluorosulfonate) and OCIO3 (perchlorate).
- OS ⁇ 2-C ⁇ H -CH3 p-toluenes
- n is 2.
- the compounds of formula (I) are in the form of a (R) -enantiomer.
- Y represents a leaving group, it is preferably halogen, more preferably chlorine.
- Compounds of formula QT wherein Y is hydroxy or chlorine are preferred.
- Compounds of formula (I) can be in free form or in salt form. In that case, dihydrochloride and dihydrobromide salts are preferred. Most preferred are dihydrochloride salts.
- Compounds of formula (I) can be in the form of a solvate, which is included in the scope of the present invention.
- Such solvates include for example hydrates, alkoxide and the like.
- Compounds of formula (I) are very stable and can be used as synthesis intermediates.
- Compounds of formula (I) may be obtained by resolution of the corresponding racemic mixture which may be prepared as described in GB 2,225,320.
- Enantiomerically pure compounds of formula (I) can be produced using industrial chiral chromatographic separation by means of commercially available chiral stationary phases. This separation can more particularly be performed using chromatographic columns sold by DAICEL Company under the trademark CHIRALPAK AD, CHIRALPAK AS and CHIRALPAK OD. Preferred are CHIRALPAK AD columns.
- the process can be carried out using batch, MCC (Multi Column Chromatography) or SMB (Simulated Moving Bed) technologies.
- the process is particularly efficient when mobile phases (eluents) such as alcohol, i.e. methanol, or mixtures of alcohols with alkanes are used.
- mobile phases eluents
- the preferred alkanes are hexane, isohexane and heptane. More preferred is heptane.
- the preferred alcohols are propanol, isopropanol, ethanol and methanol. More preferred alcohols are ethanol and methanol.
- the preferred mixtures are: 5 % to 50 % of isopropanol in hexane or in heptane, 5 % to 95 % of ethanol in hexane or in heptane, 1 % to 10 % of methanol in isohexane, and 0 % to 10 % of methanol, ethanol or isopropanol in acetonitrile.
- Compounds of formula (I) may also be obtained by reaction of enantiomerically pure l-[(4-chlorophenyl)phenylmethyl]piperazine, obtainable by the method described in GB 2,225,321, with a haloalkane.
- the present invention relates to the use of compounds of general formula (I) as synthesis intermediates, especially for the preparation of pharmaceutically active compounds.
- compounds of formula (I) are used for the synthesis of enantiomerically pure cetirizine derivatives.
- a further aspect of this invention is a process for the preparation of compounds of general formula (II) as well as pharmaceutically acceptable salts thereof,
- Z 1 represents a group of formula -OR 1 or -NR 2 R 3 , in which R 1 represents hydrogen, a hydrocarbon group or an alkali metal,
- R 2 and R 3 each independently, represent hydrogen, a hydrocarbon group or - NR 2 R 3 represents a heterocycle containing up to 7 ring members,
- W 1 is a halogen and Z 2 is as defined for Z 1 .
- hydrocarbon group is defined as including monovalent radicals containing hydrogen and carbon atoms, such as straight, branched and cyclic alkyls, alkenyls, alkynyls, aryls, alkylaryls and arylalkyls containing 1-20 carbon atoms, preferably 1 to 4 carbon atoms for non-cyclic alkyl, 6 to 10 carbon atoms for aryl and 3 to 8 carbon atoms for cycloalkyl, as well as combinations thereof.
- salts refers to salts prepared from pharmaceutically acceptable non-toxic acids or bases including inorganic acids and bases and organic acids and bases. Since the compounds of the present invention are basic, salts may be prepared from pharrnaceutlcally acceptable non-toxic acids including inorganic and organic salts.
- Suitable pharmaceutically acceptable acid addition salts for the compounds of the present invention include salts of acetic, benzenesulfonic, benzoic, camphorsulfonic, citric, ethenesulfonic, fumaric, gluconic, glutamic, hydrobromic, hydrochloric, isethionic, lactic, maleic, malic, mandelic, methanesulfonic, mucic, nitric, pantothenic, phosphoric, succinic, sulfuric, tartaric, p-toluenesulfonic acid and the like.
- Z 1 represents a group of formula -OR 1 wherein R 1 is hydrogen or a C 1 -C4-alkyl, more preferably R 1 is hydrogen or methyl, most preferably R 1 is hydrogen. According to another preferred embodiment of the above mentioned process,
- W 1 is chlorine
- Z 2 represents a group of formula -OR 1 wherein R 1 is an alkali metal, more preferably R 1 is sodium.
- the reaction of a compound of formula (I) with a compound of formula (III) is generally carried out in the presence of a chemically inert solvent and in the presence of a proton acceptor such as, for example, alkali metal hydrides, alkali metal hydroxydes, alkali metal alkoxydes or alkali metals.
- a proton acceptor such as, for example, alkali metal hydrides, alkali metal hydroxydes, alkali metal alkoxydes or alkali metals.
- Any solvent such as aliphatic and aromatic hydrocarbons, ethers, amides and alcohols of low reactivity may be used.
- the preferred solvent is THF.
- the reaction is generally carried out at a temperature of from 0°C to the reflux temperature of the reaction mixture.
- the compounds of formula (I) are used for the synthesis of enantiomerically pure compounds of general formula (TV).
- the present invention also relates to a process for the preparation of compounds of general formula (TV), as well as pharmaceutically acceptable salts thereof,
- Z 3 represents a group of formula -OR 1 ' or - ⁇ R 2 'R 3 ', in which R 1 ' represents hydrogen, a hydrocarbon group or an alkali metal, R 2' and R 3 ' , each independently, represent hydrogen, a hydrocarbon group or -NR 2 'R 3 ' represents a heterocycle containing up to 7 ring members, comprising the use of a compound of general formula (I) in the form of a (R)- enantiomer as synthesis intermediate.
- n is preferably 2.
- compounds of formula (IN) are prepared by (a) reaction of a compound of formula (V)
- P 1 represents a hydroxy protecting group and P 2 represents hydrogen or an axnine protecting group, -with the proviso that P 1 and P 2 may be linked to form a single protecting group when P 2 is not hydrogen, with a compound of formula (I) wherein Y represents a leaving group especially with a compound of formula (I) wherein Y is halogen, most preferably Cl, and (b) a deprotection step.
- protecting group refers to a substituent that is commonly employed to block or protect one or more functionalities while carrying out reactions with other functional groups on the compound.
- an "amine protecting group” is a substituent attached to an amino group that blocks or protects the amino functionality in the compound.
- a "hydroxy protecting group” refers to a substituent of a hydroxy group that blocks or protects the hydroxy functionality.
- the protecting group is preferably dialkylmethylene, more preferably isopropylene (dimethylmethylene).
- the compound of formula (V) is
- Compounds of formula (V) may be prepared by reacting a compound of formula (VI) wherein R 5 is as defined for R 4 and W 2 is halogen with a compound of formula (VII), for example under Sonogashira or related conditions (ref. : Angew. Chem. Int. Ed. 2002, 41, 4176-4211).
- reaction is generally carried out in the presence of organic or inorganic proton acceptor such as tiietj ylainine or di-isopropylamine, and solvent such as ethylacetate, DMF, MTBE (methyl terbutyl ether), i-propylacetate, toluene, water or a mixture thereof.
- organic or inorganic proton acceptor such as tiietj ylainine or di-isopropylamine
- solvent such as ethylacetate, DMF, MTBE (methyl terbutyl ether), i-propylacetate, toluene, water or a mixture thereof.
- Compounds of formula (VII) can for example be obtained by reacting a compound of formula (VIII) with the appropriate reagent to introduce the required protecting groups P 1 and P 2 .
- Compounds of formula (Vila) are preferably obtained by reacting a compound of formula (VIII) with acetone, 2,2-dimethoxypropane or methoxypropane, in the presence of acid, preferably sulfuric acid.
- the deprotection of the compound obtained from the reaction of compound (V) with compound (I) can be carried out under acidic or basic conditions depending on the nature of the protecting group.
- this process comprises the steps of:
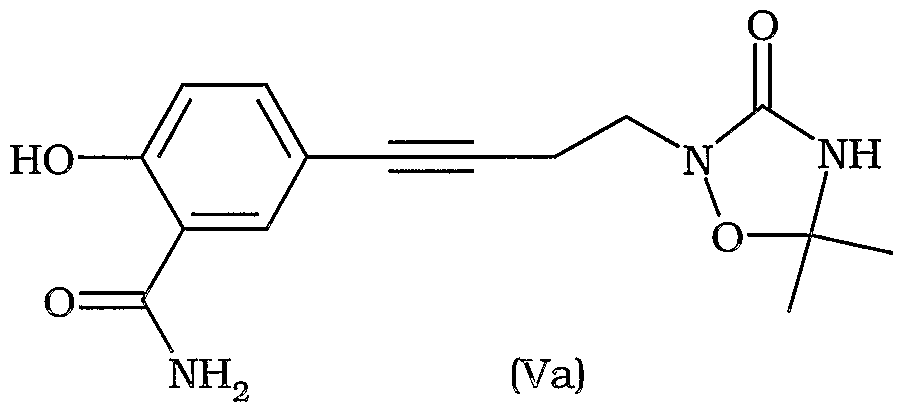
- R5 is preferably H.
- the present invention therefore also relates to synthesis intermediates of formulae (Va), (Vila) and (IX).
- Reaction between 3-butyn-l-yl methanesulfonate ester and an excess of hydroxylamine is carried out in aqueous solution in presence of methyl alcohol.
- methyl alcohol is evaporated under vacuum.
- the solution is washed with toluene.
- the aqueous phase is then washed with ethyl acetate to extract N ⁇ (3-butyn-l-yl) hydroxylamine.
- the organic phases containing N-(3-butyn-l-yl) hydroxylarnine are collected in order to be concentrated and then directly used into the next step.
- EtOAc solution of N-(3-butyn-l-yl) hydroxylamine obtained from the previous step is directly used in the next step.
- potassium cyanate previously dissolved in demineralized water.
- HC1 37% aqueous solution
- the reaction medium is then heated to 20 °C and after decantation, the two phases are separated.
- Sodium chloride is added to the aqueous phase under stirring to saturation and the aqueous phase is washed with EtOAc. All the EtOAc phases are collected and evaporated under vacuum to dryness to give crude N- (3-butyn-l-yl) N-hydroxyurea (compound VIII).
- a rnixture of compound Va, compound lc free base and K2CO3 are stirred in isopropanol or in pentyl alcohol at a temperature between 30 °C and 40 °C.
- the reaction mixture is cooled down to room temperature and filtered.
- the filter cake is washed with isopropanol or pentyl alcohol.
- the deprotection step is carried out at room temperature, in a rnixture of HOAc, H 2 SO 4 and water. As soon as the reaction is complete, the solvent is evaporated. The residual oil is dissolved in EtOAc and the solution is washed with saturated aqueous NaHC0 3 solution. The EtOAc layer is then washed twice with saturated NaCl aqueous solution. The solution is concentrated (half of the EtOAc is evaporated) using a hot water bath. The solution is stirred and allowed to cool to room temperature, and then cooled down to 0 °C. The solid is filtered, washed with EtOAc and dried to give compound 4a as a white to off-white solid (70 % yield, > 99 % purity). Purity : > 96 % ee.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Abstract
Description


Claims




Priority Applications (5)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| CA002514145A CA2514145A1 (en) | 2003-01-23 | 2004-01-20 | Piperazine derivatives and their use as synthesis intermediates |
| EP04703367A EP1590323A2 (en) | 2003-01-23 | 2004-01-20 | Piperazine derivatives and their use as synthesis intermediates |
| US10/548,709 US7381821B2 (en) | 2003-01-23 | 2004-01-20 | Piperazine derivatives and their use as synthesis intermediates |
| AU2004205494A AU2004205494B2 (en) | 2003-01-23 | 2004-01-20 | Piperazine derivatives and their use as synthesis intermediates |
| NO20053910A NO330989B1 (en) | 2003-01-23 | 2005-08-22 | New piperazine derivatives, processes for their preparation and their use as synthetic intermediates. |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| EP03001565 | 2003-01-23 | ||
| EP03001565.5 | 2003-01-23 |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| WO2004065360A2 true WO2004065360A2 (en) | 2004-08-05 |
| WO2004065360A3 WO2004065360A3 (en) | 2004-11-11 |
Family
ID=32748775
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/EP2004/000399 Ceased WO2004065360A2 (en) | 2003-01-23 | 2004-01-20 | Piperazine derivatives and their use as synthesis intermediates |
Country Status (8)
| Country | Link |
|---|---|
| US (1) | US7381821B2 (en) |
| EP (1) | EP1590323A2 (en) |
| CN (1) | CN100378086C (en) |
| AU (1) | AU2004205494B2 (en) |
| CA (1) | CA2514145A1 (en) |
| NO (1) | NO330989B1 (en) |
| PL (1) | PL378027A1 (en) |
| WO (1) | WO2004065360A2 (en) |
Cited By (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN101128446B (en) * | 2005-03-03 | 2010-12-08 | Ucb法奇姆股份有限公司 | Pyroglutamate salts and their use in optical resolution of intermediates for the synthesis of dextrocetirizine and levocetirizine |
| US20110172425A1 (en) * | 2008-09-17 | 2011-07-14 | Calyx Chemicals And Pharmaceuticals Pvt. Ltd. | Novel water based process for the preparation of substituted diphenylmethyl piperazines |
| US8049011B2 (en) | 2007-03-12 | 2011-11-01 | Krka, Tovarna Zdravil, D.D., Novo Mesto | Process for the preparation of levocetirizine and intermediates thereof |
Families Citing this family (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP1590323A2 (en) | 2003-01-23 | 2005-11-02 | Ucb Farchim, S.A. | Piperazine derivatives and their use as synthesis intermediates |
| EP2175856A4 (en) * | 2007-08-15 | 2011-08-24 | Chemagis Ltd | Novel process for preparing highly pure levocetirizine and salts thereof |
| CN103497166A (en) * | 2013-09-27 | 2014-01-08 | 盐城格瑞茵化工有限公司 | Synthesis method of cetirizine hydrochloride intermediate |
Family Cites Families (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| GB8827391D0 (en) * | 1988-11-23 | 1988-12-29 | Ucb Sa | Process for preparation of 2-(2-(4-((4-chlorophenyl)phenylmethyl)-1-pipera-zinyl)ethoxy)-acetic acid & its dihydrochloride |
| GB8827390D0 (en) * | 1988-11-23 | 1988-12-29 | Ucb Sa | Process for preparation of 2-(2-(4-((4-chlorophenyl)phenylmethyl)-1-pipera-zinyl)ethoxy)-acetic acid & its dihydrochloride |
| WO1993002062A1 (en) | 1991-07-19 | 1993-02-04 | Zeria Pharmaceutical Co., Ltd. | Piperazine derivative and drug containing the same |
| PE20001566A1 (en) | 1999-03-26 | 2001-02-05 | Ucb Sa | 1,4-SUBSTITUTED PIPERAZINES, 1,4-SUBSTITUTED PIPERIDINES AND 1-SUBSTITUTED 4-ALKYLIDENYL PIPERIDINES |
| GB0009479D0 (en) * | 2000-04-17 | 2000-06-07 | Cipla Limited | Antihistaminic compounds |
| EP1590323A2 (en) | 2003-01-23 | 2005-11-02 | Ucb Farchim, S.A. | Piperazine derivatives and their use as synthesis intermediates |
-
2004
- 2004-01-20 EP EP04703367A patent/EP1590323A2/en not_active Ceased
- 2004-01-20 PL PL378027A patent/PL378027A1/en not_active Application Discontinuation
- 2004-01-20 CN CNB2004800026561A patent/CN100378086C/en not_active Expired - Fee Related
- 2004-01-20 AU AU2004205494A patent/AU2004205494B2/en not_active Ceased
- 2004-01-20 CA CA002514145A patent/CA2514145A1/en not_active Abandoned
- 2004-01-20 US US10/548,709 patent/US7381821B2/en not_active Expired - Fee Related
- 2004-01-20 WO PCT/EP2004/000399 patent/WO2004065360A2/en not_active Ceased
-
2005
- 2005-08-22 NO NO20053910A patent/NO330989B1/en not_active IP Right Cessation
Non-Patent Citations (1)
| Title |
|---|
| See references of EP1590323A2 * |
Cited By (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN101128446B (en) * | 2005-03-03 | 2010-12-08 | Ucb法奇姆股份有限公司 | Pyroglutamate salts and their use in optical resolution of intermediates for the synthesis of dextrocetirizine and levocetirizine |
| US8049011B2 (en) | 2007-03-12 | 2011-11-01 | Krka, Tovarna Zdravil, D.D., Novo Mesto | Process for the preparation of levocetirizine and intermediates thereof |
| US20110172425A1 (en) * | 2008-09-17 | 2011-07-14 | Calyx Chemicals And Pharmaceuticals Pvt. Ltd. | Novel water based process for the preparation of substituted diphenylmethyl piperazines |
Also Published As
| Publication number | Publication date |
|---|---|
| EP1590323A2 (en) | 2005-11-02 |
| US20060183903A1 (en) | 2006-08-17 |
| US7381821B2 (en) | 2008-06-03 |
| CN100378086C (en) | 2008-04-02 |
| CN1742002A (en) | 2006-03-01 |
| AU2004205494A1 (en) | 2004-08-05 |
| WO2004065360A3 (en) | 2004-11-11 |
| CA2514145A1 (en) | 2004-08-05 |
| NO330989B1 (en) | 2011-08-29 |
| NO20053910D0 (en) | 2005-08-22 |
| PL378027A1 (en) | 2006-02-20 |
| NO20053910L (en) | 2005-10-21 |
| AU2004205494B2 (en) | 2009-04-30 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP0529842B1 (en) | Production of fluoxetine and new intermediates | |
| US8518936B2 (en) | Method for preparing acid addition salts of polyacidic basic compounds | |
| JP2017503843A (en) | N-[(3-aminooxetane-3-yl) methyl] -2- (1,1-dioxo-3,5-dihydro-1,4-benzothiazepin-4-yl) -6-methyl-quinazoline- Process for the preparation of 4-amine | |
| IE64038B1 (en) | Tertiary alkyl functionalized piperazine derivatives | |
| KR20060126965A (en) | Process for preparing O-carbamoyl compound in the presence of active amine group | |
| KR101597328B1 (en) | Phenyl piperazine compounds, pharmaceutical composition comprising the same, and use thereof | |
| EP3891133B1 (en) | Improved process for preparing ozanimod | |
| US10584100B2 (en) | Solid state forms of netarsudil mesylate | |
| US7381821B2 (en) | Piperazine derivatives and their use as synthesis intermediates | |
| EP2062881B1 (en) | Process for making N-(diphenylmethyl)piperazines | |
| GB2142021A (en) | 1 4-dihydropyridine derivatives their preparation and pharmaceutical compositions containing them | |
| WO2024191675A1 (en) | Methods for the preparation of sstr4 agonists and salts thereof | |
| EP4317141A1 (en) | Process and intermediates for the preparation of viloxazine and other 2-substituted morpholine derivatives | |
| US7875750B2 (en) | Method of obtaining 2-amino-6-alkyl-amino-4,5,6,7-tetrahydrobenzothiazoles | |
| KR100928776B1 (en) | (R) -1-[(4-chlorophenyl) phenylmethyl] piperazine or a salt thereof | |
| KR100503443B1 (en) | Processes for preparing an optically active cetirizine or its salt | |
| CZ218999A3 (en) | A 4- (1-pierazinyl) benzoic acid derivative, a process for its preparation, and a pharmaceutical composition thereof | |
| KR100998067B1 (en) | Bis (1-[(4-chlorophenyl) phenylmethyl] piperazin) -2,3-dibenzoyl tartaric acid novel intermediate salt and optically pure 1-[(4-chlorophenyl) phenylmethyl] piperazin using the same How to separate | |
| BE898219A (en) | New amine derivatives, their salts, process for preparing them, and anti-ulcer agent containing them. | |
| CN119569587A (en) | Preparation method of alkoxy diethyl amine | |
| US20070270483A1 (en) | Process for the preparation of aryloxypropylamines | |
| EA046640B1 (en) | METHOD AND INTERMEDIATE COMPOUND FOR OBTAINING OXETAN-2-YLMETHANAMINE | |
| EA049781B1 (en) | METHOD FOR PRODUCING AVAPRITINIB AND ITS INTERMEDIATES | |
| FR2784990A1 (en) | Preparation of 4-(2-amino-thiazol-4-yl)-benzamidine derivatives, useful as antithrombotic agents, comprises reacting nitrile compounds with hydroxylamine then reducing the obtained intermediate | |
| JPH0772183B2 (en) | Method for producing benzothiazine derivative |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| AK | Designated states |
Kind code of ref document: A2 Designated state(s): AE AG AL AM AT AU AZ BA BB BG BR BW BY BZ CA CH CN CO CR CU CZ DE DK DM DZ EC EE EG ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX MZ NA NI NO NZ OM PG PH PL PT RO RU SC SD SE SG SK SL SY TJ TM TN TR TT TZ UA UG US UZ VC VN YU ZA ZM ZW |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A2 Designated state(s): BW GH GM KE LS MW MZ SD SL SZ TZ UG ZM ZW AM AZ BY KG KZ MD RU TJ TM AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IT LU MC NL PT RO SE SI SK TR BF BJ CF CG CI CM GA GN GQ GW ML MR NE SN TD TG |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 2004703367 Country of ref document: EP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 3098/DELNP/2005 Country of ref document: IN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 378027 Country of ref document: PL Ref document number: 20048026561 Country of ref document: CN Ref document number: 2004205494 Country of ref document: AU Ref document number: 2514145 Country of ref document: CA |
|
| WWP | Wipo information: published in national office |
Ref document number: 2004205494 Country of ref document: AU |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2006183903 Country of ref document: US Ref document number: 10548709 Country of ref document: US |
|
| WWP | Wipo information: published in national office |
Ref document number: 2004703367 Country of ref document: EP |
|
| WWP | Wipo information: published in national office |
Ref document number: 10548709 Country of ref document: US |