EP2712858A1 - Process for producing thionocarboxylic acid ester - Google Patents
Process for producing thionocarboxylic acid ester Download PDFInfo
- Publication number
- EP2712858A1 EP2712858A1 EP13184535.6A EP13184535A EP2712858A1 EP 2712858 A1 EP2712858 A1 EP 2712858A1 EP 13184535 A EP13184535 A EP 13184535A EP 2712858 A1 EP2712858 A1 EP 2712858A1
- Authority
- EP
- European Patent Office
- Prior art keywords
- formula
- optionally substituted
- compounds
- compound represented
- represented
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Granted
Links
- 0 *c1ccccc1 Chemical compound *c1ccccc1 0.000 description 3
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C327/00—Thiocarboxylic acids
- C07C327/20—Esters of monothiocarboxylic acids
- C07C327/26—Esters of monothiocarboxylic acids having carbon atoms of esterified thiocarboxyl groups bound to carbon atoms of six-membered aromatic rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C315/00—Preparation of sulfones; Preparation of sulfoxides
- C07C315/04—Preparation of sulfones; Preparation of sulfoxides by reactions not involving the formation of sulfone or sulfoxide groups
Definitions
- the present invention relates to a process for producing an ester of aryl-containing thionocarboxylic acid that is useful as a liquid crystal material, etc.
- the aryl ester of arylthionocarboxylic acid represented by the formula R 1 C (S)OR 2 , wherein R 1 and R 2 independently represent an optionally substituted aryl group, is a compound that is useful as a liquid crystal material, etc.
- the methods disclosed in NPL 1, 2, 4, 7, etc. are low-yield; in particular, the method disclosed in NPL 4 requires a high temperature of 100°C or more for the reaction, and also has a drawback of generating a large amount of by-products; therefore, these methods cannot be regarded as efficient methods.
- NPL 3 and 6 use hydrogen sulfide, which is so toxic that its handling requires specialized equipment; therefore, these methods are unsuitable as industrial processes for production. Further, the method disclosed in NPL 5 cannot be regarded as a comprehensively superior method due to a low synthesis yield of the starting thionocarboxylic acid chloride.
- the method disclosed in NPL 8 comprises a large number of steps, including synthesizing a thioacyl disulfide from a dithiocarboxylic acid derivative and then reacting the thioacyl disulfide with an alcoholate; therefore, this method cannot be regarded as a convenient method.
- a primary object of the invention is to provide a relatively convenient and industrially advantageous process for producing, in a high yield, an arylthionocarboxylic acid ester, which is useful as a liquid crystal material, etc.
- a primary object of the invention is to provide a relatively convenient and industrially advantageous process for producing, in a high yield, an arylthionocarboxylic acid ester, which is useful as a liquid crystal material, etc.
- the present inventors conducted extensive research. As a result, the inventors found that when a halogenated thiocarbonyl compound is used as a starting material and reacted with a Grignard compound having an aryl group or a cycloalkyl group, the desired arylthionocarboxylic acid ester can be produced in a high yield under mild reaction conditions using relatively safe starting materials.
- the present invention has been accomplished based on this finding. More specifically, the present invention provides the following processes for producing an arylthionocarboxylic acid ester.
- Item 1 A process for producing an arylthionocarboxylic acid ester represented by Formula (3): wherein Ar is an optionally substituted aryl group and R is an optionally substituted aryl group or an optionally substituted cycloalkyl group; the process comprising reacting a halogenated thiocarbonyl compound represented by Formula (1): wherein Ar is as defined above and X is a halogen atom, with a Grignard compound represented by Formula (2): R-MgY wherein R is as defined above and Y is a halogen atom.
- Item 2 The process according to Item 1, wherein the reaction is performed in an aprotic organic solvent.
- Item 3 The process according to Item 1 or 2, wherein the halogenated thiocarbonyl compound represented by Formula (1) is reacted with the Grignard compound represented by Formula (2) in the presence of at least one component selected from the group consisting of trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, and divalent cobalt compounds.
- Item 4 A process for producing an arylthionocarboxylic acid ester represented by Formula (3): wherein Ar is an optionally substituted aryl group and R is an optionally substituted aryl group or an optionally substituted cycloalkyl group; the process comprising reacting a halide represented by Formula (4): R-Y wherein R is as defined above and Y is a halogen atom, with a magnesium compound, and then reacting the reaction product with a halogenated thiocarbonyl compound represented by Formula (1): wherein Ar is as defined above and X is a halogen atom.
- the process of the present invention for producing an arylthionocarboxylic acid ester comprises reacting a halogenated thiocarbonyl compound represented by Formula (1): wherein Ar is an optionally substituted aryl group and X is a halogen atom, with a Grignard compound represented by Formula (2): R-MgY wherein R is an optionally substituted aryl group or an optionally substituted cycloalkyl group and Y is a halogen atom.
- This process can produce an arylthionocarboxylic acid ester represented by Formula (3): wherein Ar and R are as defined above, in a high yield under relatively mild conditions.
- Ar is an optionally substituted aryl group.
- optionally substituted aryl groups include optionally substituted phenyl, optionally substituted naphthyl, and the like.
- substituent on the aryl group there is no particular limitation on the substituent on the aryl group, insofar as the substituent is inert to the reaction with a Grignard compound represented by Formula (2): R-MgY described below.
- substituents include alkyl, fluoroalkyl, alkoxy, fluoroalkoxy, cyano, aryl, halogen, and the like.
- alkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms
- fluoroalkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine.
- alkoxy groups include linear or branched lower alkoxy groups having about 1 to 5 carbon atoms.
- fluoroalkoxy groups include linear or branched lower alkoxy groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine.
- aryl groups as substituents include phenyl, naphthyl, and the like. The substituent aryl groups may further contain a substituent that is inert to the reaction.
- halogen atoms include fluorine, chlorine, bromine, and iodine.
- the halogen atom represented by X may be, for example, fluorine, chlorine, bromine, or iodine. Chlorine is particularly preferable.
- the cycloalkyl group of the optionally substituted cycloalkyl group represented by R may be, for example, a cycloalkyl group having about 3 to 8 carbon atoms, and preferably about 5 to 7 carbon atoms.
- Examples of such cycloalkyl groups include cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl, cycloheptyl, cyclooctyl, and the like.
- alkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms; and examples of fluoroalkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine.
- fluoroalkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine.
- cycloaryl groups as substituents include cycloalkyl groups having about 3 to 8 carbon atoms.
- the substituent cycloaryl groups may further contain a substituent that is inert to the reaction, such as alkyl.
- the magnesium compound that can be used in this reaction is not particularly limited insofar as the compound is reactive with a halide represented by Formula (4) to produce a Grignard compound.
- Specific examples of such magnesium compounds include metal magnesium, i-PrMgCl (isopropylmagnesium chloride), i-PrMgBr (isopropylmagnesium bromide), and an i-PrBu 2 MgLi ate complex which can be produced from n-BuLi and i-PrMgCl or i-PrMgBr.
- the amount of the magnesium compound may be about 1.0 to 10.0 mol, and preferably about 1.0 to 2.0 mol, per mol of the halide represented by Formula (4): R-Y.
- the reaction of a halide represented by Formula (4): R-Y with a magnesium compound is preferably performed in an aprotic organic solvent inert to the reaction.
- aprotic organic solvents include tetrahydrofuran, diethyl ether, 2-methyltetrahydrofuran, dibutyl ether, tert -butyl methyl ether, cyclopentyl methyl ether, dimethoxyethane, dioxane, benzene, toluene, xylene, pentane, hexane, heptane, and the like.
- Such aprotic organic solvents may be used as a mixture of two or more, if necessary.
- concentrations of the starting materials in the reaction solvent there is no particular limitation on the concentrations of the starting materials in the reaction solvent.
- concentration of the halide represented by Formula (4): R-Y may be about 0.5 to 10.0 mol/L.
- the temperature at which the reaction of a halide represented by Formula (4): R-Y with a magnesium compound is performed is preferably about 0 to 100°C, and more preferably about 20 to 70°C.
- the pressure during the reaction is not particularly limited, and can typically be performed under atmospheric pressure.
- the atmosphere during the reaction is not particularly limited.
- the reaction is preferably performed in an inert gas atmosphere, such as a nitrogen atmosphere or an argon atmosphere.
- the reaction time may be about 30 minutes to about 10 hours.
- the yield of the Grignard compound represented by Formula (2): R-MgY can be increased by reacting a halide represented by Formula (4): R-Y with a magnesium compound in the presence of LiCl, iodine, dibromoethane, or the like ingredients.
- the total amount of these ingredients is preferably about 1.0 to 5.0 mol, and more preferably about 1.0 to 1.2 mol, per mol of the halide represented by Formula (4): R-Y.
- the above process can produce a Grignard compound represented by Formula (2): R-MgY wherein R is an optionally substituted aryl group or an optionally substituted cycloalkyl group and Y is a halogen atom.
- the desired arylthionocarboxylic acid ester of the present invention which is represented by Formula (3): wherein Ar is an optionally substituted aryl group and R is an optionally substituted aryl group or an optionally substituted cycloalkyl group, can be obtained by reacting a halogenated thiocarbonyl compound represented by Formula (1): wherein Ar is as defined above and X is a halogen atom, with a Grignard compound represented by Formula (2): R-MgY wherein R is as defined above and Y is a halogen atom.
- the amount of the Grignard compound represented by Formula (2): R-MgY wherein R and Y are as defined above may be about 0.8 to 2.0 mol, and preferably about 0.9 to 1.2 mol, per mol of the halogenated thiocarbonyl compound represented by Formula (1).
- the reaction of a halogenated thiocarbonyl compound represented by Formula (1) with a Grignard compound represented by Formula (2): R-MgY is preferably performed in an aprotic organic solvent.
- aprotic organic solvent examples include the same as the aforementioned examples of solvents that can be used in the reaction of a halide represented by Formula (4): R-Y with a magnesium compound.
- the concentrations of the starting materials in the reaction solvent may be about 0.1 mol/L to 3.0 mol/L.
- the temperature at which the reaction of a halogenated thiocarbonyl compound with a Grignard compound is performed is such that when using a Grignard compound wherein R is an optionally substituted aryl group, the temperature is preferably about -78 to 25°C, and more preferably about -50 to 0°C: and when using a Grignard compound wherein R is an optionally substituted cycloalkyl group, the temperature is preferably about -78 to 0°C, and more preferably about -78 to -30°C.
- the yield of the desired arylthionocarboxylic acid ester represented by Formula (3) can be increased particularly by reacting a halogenated thiocarbonyl compound represented by Formula (1) with a Grignard compound represented by Formula (2): R-MgY in the presence of a catalyst.
- a catalyst include trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, divalent cobalt compounds, and the like. Such compounds can be used singly, or as a mixture of two or more.
- trivalent iron compounds include tris(2,4-pentanedionato)iron(III), ferric chloride, and the like.
- Examples of monovalent or divalent copper compounds include copper halides such as copper(I) chloride, copper(I) bromide, and copper(I) iodide; copper cyanide (CuCN), cupric chloride, dilithium tetrachlorocuprate(II)(Li 2 C U Cl 4 ), and the like.
- Examples of divalent nickel compounds include nickel chloride and the like.
- Examples of divalent zinc compounds include zinc chloride and the like.
- Examples of divalent cobalt compounds include cobalt chloride and the like.
- Such a catalyst is preferably used in an amount of about 0.01 to 2.00 mol, and more preferably 0.01 to 0.10 mol, per mol of the halogenated thiocarbonyl compound represented by Formula (1).
- the above process can produce an arylthionocarboxylic acid ester represented by Formula (3): in a high yield.
- reaction conditions in the step of reacting a halide represented by Formula (4) with a magnesium compound and in the step of reacting the reaction product with a halogenated thiocarbonyl compound represented by Formula (1) in the continuous process may be the same as the aforementioned conditions in each reaction step.
- the arylthionocarboxylic acid ester obtained by this process can be purified and collected by a known method, such as silica gel column chromatography or recrystallization.
- the desired arylthionocarboxylic acid ester can be obtained in a high yield by using relatively inexpensive and highly safe starting materials under mild reaction conditions.
- phenyl chlorothionoformate (11.0 g, 64.0 mmol), tris(2,4-pentanedionato)iron(III) (Fe(acac) 3 ) (0.7 g, 1.9 mmol), and THF (50.0 mL) were added in a nitrogen atmosphere to a 200 ml three-necked flask equipped with a stirring bar and a thermometer. After the mixture was cooled to -30°C, the Grignard compound obtained by the above process was added, and the resulting mixture was stirred for 2 hours to complete the reaction.
- Tables 1 and 2 show the kinds and yields of the obtained products.
- the each Grignard compound shown in the Compound (2) column of the table was obtained by reacting magnesium with a halide serving as precursor of the each Grignard compound in the same manner as in Example 1.
- the numerical values shown in the Catalyst column of the table are the amounts of catalyst expressed in mol% relative to Compound (1).
- Table 1 Example Compound (1) Catalyst Compound (2) Solvent Temp.
- phenyl chlorothionoformate (11.0 g, 64 mmol), a 0.1 M tetrahydrofuran solution of dilithium tetrachlorocuprate(II)(19 mL, 1.9 mmol), and THF (10 mL) were added in a nitrogen atmosphere to a 200 ml three-necked flask equipped with a stirring bar and a thermometer. After the mixture was cooled to -30°C, the Grignard compound obtained by the above process was added, and the resulting mixture was stirred for 2 hours to complete the reaction. This mixture was returned to room temperature, and extracted with toluene/hydrochloric acid. The organic layer was evaporated under reduced pressure. The obtained solid was purified by silica gel column chromatography to give cyclohexyl thionobenzoate as a yellow solid (10.1 g, 46 mmol). The yield was 72%.
- Example 3 shows the kinds and yields of the obtained products.
- the each Grignard compound shown in the Compound (2) column of the table was obtained by reacting magnesium with a halide serving as precursor of each Grignard compound in the same manner as in Example 20. Further, the numerical values shown in the Catalyst column of the table are the amounts of catalyst expressed in mol% relative to Compound (1).
- Table 3 Example Compound (1) Catalyst Compound (2) Solvent Temp.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
Abstract

Description
- The present invention relates to a process for producing an ester of aryl-containing thionocarboxylic acid that is useful as a liquid crystal material, etc.
- The aryl ester of arylthionocarboxylic acid represented by the formula R1C=(S)OR2, wherein R1 and R2 independently represent an optionally substituted aryl group, is a compound that is useful as a liquid crystal material, etc.
- Various methods are known as methods for producing thionocarboxylic acid esters having this structure. Examples of such methods include a method comprising reacting a chlorothionoformic acid ester with an aromatic compound (Non-patent Literature (NPL) 1); a method comprising treating a carbanion with a dithiocarbonic acid ester or a thionocarbonic acid ester (Non-patent Literature (NPL) 2); a method comprising reacting an orthoester with hydrogen sulfide (Non-patent Literature (NPL) 3); a method comprising reacting a carboxylic acid ester with phosphorus pentasulfide or Lawesson's reagent (Non-patent Literature (NPL) 4); a method comprising reacting a thionocarboxylic acid chloride with an alcohol or phenol (Non-patent Literature (NPL) 5); a method comprising reacting a nitrile with an alcohol and then reacting the resulting product with hydrogen sulfide (Non-patent Literature (NPL) 6); a method comprising treating a methylated aromatic compound with sulfur and an alcohol (Non-patent Literature (NPL) 7); and a method comprising reacting a thioacyl disulfide with an alcoholate (Non-patent Literature (NPL) 8).
- However, among these methods, the methods disclosed in NPL 1, 2, 4, 7, etc., are low-yield; in particular, the method disclosed in NPL 4 requires a high temperature of 100°C or more for the reaction, and also has a drawback of generating a large amount of by-products; therefore, these methods cannot be regarded as efficient methods.
- The methods disclosed in NPL 3 and 6 use hydrogen sulfide, which is so toxic that its handling requires specialized equipment; therefore, these methods are unsuitable as industrial processes for production. Further, the method disclosed in NPL 5 cannot be regarded as a comprehensively superior method due to a low synthesis yield of the starting thionocarboxylic acid chloride. The method disclosed in NPL 8 comprises a large number of steps, including synthesizing a thioacyl disulfide from a dithiocarboxylic acid derivative and then reacting the thioacyl disulfide with an alcoholate; therefore, this method cannot be regarded as a convenient method.
- Non-patent Literature
- NPL 1:
- H. Viola, et al., Chem. Ber., 101, 3517 (1968)
- NPL 2:
- Liebigs Ann. Chem., 1973, 1637
- NPL 3:
- A. Ohno et al., Tetrahedron Lett., 1968, 2083
- NPL 4:
- Synthesis, 1973, 149; Bull. Chem. Soc. Belg., 87, 293 (1987)
- NPL 5:
- S. Scheithauer et al., Chem. Ber., 98, 838 (1965)
- NPL 6:
- Liebigs Ann. Chem., 1974, 671
- NPL 7:
- Z. Chem., 6, 108 (1966)
- NPL 8:
- K.A. Latif et al., Tetrahedron, 26, 4247 (1970)
- The present invention was made in view of the above-mentioned problems in the prior art. A primary object of the invention is to provide a relatively convenient and industrially advantageous process for producing, in a high yield, an arylthionocarboxylic acid ester, which is useful as a liquid crystal material, etc.
- The present invention was made in view of the above-mentioned problems in the prior art. A primary object of the invention is to provide a relatively convenient and industrially advantageous process for producing, in a high yield, an arylthionocarboxylic acid ester, which is useful as a liquid crystal material, etc.
- To achieve the above object, the present inventors conducted extensive research. As a result, the inventors found that when a halogenated thiocarbonyl compound is used as a starting material and reacted with a Grignard compound having an aryl group or a cycloalkyl group, the desired arylthionocarboxylic acid ester can be produced in a high yield under mild reaction conditions using relatively safe starting materials. The present invention has been accomplished based on this finding. More specifically, the present invention provides the following processes for producing an arylthionocarboxylic acid ester.
- Item 1. A process for producing an arylthionocarboxylic acid ester represented by Formula (3):

the process comprising reacting a halogenated thiocarbonyl compound represented by Formula (1):
- Item 2. The process according to Item 1, wherein the reaction is performed in an aprotic organic solvent.
- Item 3. The process according to Item 1 or 2, wherein the halogenated thiocarbonyl compound represented by Formula (1) is reacted with the Grignard compound represented by Formula (2) in the presence of at least one component selected from the group consisting of trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, and divalent cobalt compounds.
- Item 4. A process for producing an arylthionocarboxylic acid ester represented by Formula (3):

the process comprising reacting a halide represented by Formula (4): R-Y wherein R is as defined above and Y is a halogen atom, with a magnesium compound, and then reacting the reaction product with a halogenated thiocarbonyl compound represented by Formula (1):
- Item 5. The process according to Item 4, wherein the reaction with the halogenated thiocarbonyl compound represented by Formula (1) is performed in the presence of at least one component selected from the group consisting of trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, and divalent cobalt compounds.
- The process of the present invention for producing an arylthionocarboxylic acid ester is explained in more detail below.
- The process of the present invention for producing an arylthionocarboxylic acid ester comprises reacting a halogenated thiocarbonyl compound represented by Formula (1):


- First, the starting compounds used in the production process of the present invention are explained in more detail.
- In the starting halogenated thiocarbonyl compound represented by Formula (1), Ar is an optionally substituted aryl group. Examples of optionally substituted aryl groups include optionally substituted phenyl, optionally substituted naphthyl, and the like.
- There is no particular limitation on the substituent on the aryl group, insofar as the substituent is inert to the reaction with a Grignard compound represented by Formula (2): R-MgY described below. Specific examples of such substituents include alkyl, fluoroalkyl, alkoxy, fluoroalkoxy, cyano, aryl, halogen, and the like. Among these, examples of alkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms; and examples of fluoroalkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine. Examples of alkoxy groups include linear or branched lower alkoxy groups having about 1 to 5 carbon atoms. Examples of fluoroalkoxy groups include linear or branched lower alkoxy groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine. Examples of aryl groups as substituents include phenyl, naphthyl, and the like. The substituent aryl groups may further contain a substituent that is inert to the reaction. Examples of halogen atoms include fluorine, chlorine, bromine, and iodine.
- In the above Formula (1), the halogen atom represented by X may be, for example, fluorine, chlorine, bromine, or iodine. Chlorine is particularly preferable.
- In the Grignard compound represented by Formula (2): R-MgY wherein R is an optionally substituted aryl group or an optionally substituted cycloaryl group and Y is a halogen atom, the optionally substituted aryl group represented by R may be, for example, an optionally substituted phenyl group or an optionally substituted naphthyl group. The substituent on the aryl group may be any substituent that is inert to the reaction with a halogenated thiocarbonyl compound represented by Formula (1), and that is also inert to the reaction for producing a Grignard compound represented by Formula (2): R-MgY from a halide represented by Formula (4): R-Y described below. Specific examples of such substituents may be the same as the aforementioned examples of substituents on the aryl group represented by Ar.
- The cycloalkyl group of the optionally substituted cycloalkyl group represented by R may be, for example, a cycloalkyl group having about 3 to 8 carbon atoms, and preferably about 5 to 7 carbon atoms. Examples of such cycloalkyl groups include cyclopropyl, cyclobutyl, cyclopentyl, cyclohexyl, cycloheptyl, cyclooctyl, and the like. The substituent on the cycloalkyl group may be any substituent that is inert to the reaction with a halogenated thiocarbonyl compound represented by Formula (1), and that is also inert to the reaction for producing a Grignard compound represented by Formula (2): R-MgY from a halide represented by Formula (4): R-Y described below. Specific examples of such substituents include alkyl, cycloalkyl, fluoroalkyl, and the like. Among such substituents, examples of alkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms; and examples of fluoroalkyl groups include linear or branched lower alkyl groups having about 1 to 5 carbon atoms in which hydrogen atoms are partially or completely replaced by fluorine. Examples of cycloaryl groups as substituents include cycloalkyl groups having about 3 to 8 carbon atoms. The substituent cycloaryl groups may further contain a substituent that is inert to the reaction, such as alkyl.
- To obtain a Grignard compound represented by Formula (2): R-MgY, for example, a halide represented by Formula (4): R-Y may be reacted with a magnesium compound.
- The magnesium compound that can be used in this reaction is not particularly limited insofar as the compound is reactive with a halide represented by Formula (4) to produce a Grignard compound. Specific examples of such magnesium compounds include metal magnesium, i-PrMgCl (isopropylmagnesium chloride), i-PrMgBr (isopropylmagnesium bromide), and an i-PrBu2MgLi ate complex which can be produced from n-BuLi and i-PrMgCl or i-PrMgBr.
- The amount of the magnesium compound may be about 1.0 to 10.0 mol, and preferably about 1.0 to 2.0 mol, per mol of the halide represented by Formula (4): R-Y.
- Typically, the reaction of a halide represented by Formula (4): R-Y with a magnesium compound is preferably performed in an aprotic organic solvent inert to the reaction. Examples of aprotic organic solvents include tetrahydrofuran, diethyl ether, 2-methyltetrahydrofuran, dibutyl ether, tert-butyl methyl ether, cyclopentyl methyl ether, dimethoxyethane, dioxane, benzene, toluene, xylene, pentane, hexane, heptane, and the like. Such aprotic organic solvents may be used as a mixture of two or more, if necessary.
- There is no particular limitation on the concentrations of the starting materials in the reaction solvent. For example, the concentration of the halide represented by Formula (4): R-Y may be about 0.5 to 10.0 mol/L.
- The temperature at which the reaction of a halide represented by Formula (4): R-Y with a magnesium compound is performed is preferably about 0 to 100°C, and more preferably about 20 to 70°C.
- The pressure during the reaction is not particularly limited, and can typically be performed under atmospheric pressure. The atmosphere during the reaction is not particularly limited. Typically, the reaction is preferably performed in an inert gas atmosphere, such as a nitrogen atmosphere or an argon atmosphere. Typically, the reaction time may be about 30 minutes to about 10 hours.
- The yield of the Grignard compound represented by Formula (2): R-MgY can be increased by reacting a halide represented by Formula (4): R-Y with a magnesium compound in the presence of LiCl, iodine, dibromoethane, or the like ingredients. The total amount of these ingredients is preferably about 1.0 to 5.0 mol, and more preferably about 1.0 to 1.2 mol, per mol of the halide represented by Formula (4): R-Y.
- The above process can produce a Grignard compound represented by Formula (2): R-MgY wherein R is an optionally substituted aryl group or an optionally substituted cycloalkyl group and Y is a halogen atom.
- As described above, the desired arylthionocarboxylic acid ester of the present invention, which is represented by Formula (3):


- The amount of the Grignard compound represented by Formula (2): R-MgY wherein R and Y are as defined above may be about 0.8 to 2.0 mol, and preferably about 0.9 to 1.2 mol, per mol of the halogenated thiocarbonyl compound represented by Formula (1).
- The reaction of a halogenated thiocarbonyl compound represented by Formula (1) with a Grignard compound represented by Formula (2): R-MgY is preferably performed in an aprotic organic solvent. Examples of usable aprotic organic solvents may be the same as the aforementioned examples of solvents that can be used in the reaction of a halide represented by Formula (4): R-Y with a magnesium compound.
- There is no particular limitation on the concentrations of the starting materials in the reaction solvent. For example, the concentration of the halogenated thiocarbonyl compound represented by Formula (1) may be about 0.1 mol/L to 3.0 mol/L. The temperature at which the reaction of a halogenated thiocarbonyl compound with a Grignard compound is performed is such that when using a Grignard compound wherein R is an optionally substituted aryl group, the temperature is preferably about -78 to 25°C, and more preferably about -50 to 0°C: and when using a Grignard compound wherein R is an optionally substituted cycloalkyl group, the temperature is preferably about -78 to 0°C, and more preferably about -78 to -30°C.
- The pressure during the reaction is not particularly limited. The reaction can typically be performed under atmospheric pressure. The atmosphere during the reaction is not particularly limited. Typically, the reaction is preferably performed in an inert gas atmosphere, such as a nitrogen atmosphere or an argon atmosphere. Typically, the reaction time may be about 0.5 to 10.0 hours.
- In the production process of the present invention, the yield of the desired arylthionocarboxylic acid ester represented by Formula (3) can be increased particularly by reacting a halogenated thiocarbonyl compound represented by Formula (1) with a Grignard compound represented by Formula (2): R-MgY in the presence of a catalyst. Examples of usable catalysts include trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, divalent cobalt compounds, and the like. Such compounds can be used singly, or as a mixture of two or more. Specific examples of trivalent iron compounds include tris(2,4-pentanedionato)iron(III), ferric chloride, and the like. Examples of monovalent or divalent copper compounds include copper halides such as copper(I) chloride, copper(I) bromide, and copper(I) iodide; copper cyanide (CuCN), cupric chloride, dilithium tetrachlorocuprate(II)(Li2CUCl4), and the like. Examples of divalent nickel compounds include nickel chloride and the like. Examples of divalent zinc compounds include zinc chloride and the like. Examples of divalent cobalt compounds include cobalt chloride and the like.
- Such a catalyst is preferably used in an amount of about 0.01 to 2.00 mol, and more preferably 0.01 to 0.10 mol, per mol of the halogenated thiocarbonyl compound represented by Formula (1).
- The above process can produce an arylthionocarboxylic acid ester represented by Formula (3):

- The arylthionocarboxylic acid ester obtained by the above process can be purified and collected by a known method, such as silica gel column chromatography or recrystallization.
- In the present invention, instead of directly reacting a halogenated thiocarbonyl compound represented by Formula (1):


- The arylthionocarboxylic acid ester obtained by this process can be purified and collected by a known method, such as silica gel column chromatography or recrystallization.
- According to the present invention, the desired arylthionocarboxylic acid ester can be obtained in a high yield by using relatively inexpensive and highly safe starting materials under mild reaction conditions.
- The present invention is explained in more detail below with reference to Examples.
- Magnesium (1.6 g, 64.0 mmol), LiCl (2.7 g, 64.0 mmol), and tetrahydrofuran (hereinafter abbreviated as "THF") (10.0 mL) were added in a nitrogen atmosphere to a 50 ml three-necked flask equipped with a stirring bar and a thermometer. After chlorobenzene (7.2 g, 64 mmol) was added, the mixture was stirred at room temperature for 2 hours to produce a Grignard compound. Subsequently, phenyl chlorothionoformate (11.0 g, 64.0 mmol), tris(2,4-pentanedionato)iron(III) (Fe(acac)3) (0.7 g, 1.9 mmol), and THF (50.0 mL) were added in a nitrogen atmosphere to a 200 ml three-necked flask equipped with a stirring bar and a thermometer. After the mixture was cooled to -30°C, the Grignard compound obtained by the above process was added, and the resulting mixture was stirred for 2 hours to complete the reaction.
- This mixture was returned to room temperature, and extracted with toluene/hydrochloric acid. The organic layer was evaporated under reduced pressure. The obtained solid was purified by silica gel column chromatography to give phenyl thionobenzoate as a yellow solid (11.9 g, 55.5 mmol). The yield was 87%.
- Using the starting materials shown in Tables 1 and 2, the reaction was performed in the same manner as in Example 1. Tables 1 and 2 show the kinds and yields of the obtained products. The each Grignard compound shown in the Compound (2) column of the table was obtained by reacting magnesium with a halide serving as precursor of the each Grignard compound in the same manner as in Example 1. The numerical values shown in the Catalyst column of the table are the amounts of catalyst expressed in mol% relative to Compound (1).
Table 1 Example Compound (1) Catalyst Compound (2) Solvent Temp. °C Product Yield (%)* 1 
Fe(acac)3 3mol% 
THF -30°C 
87 2 
Fe(acac)3 3mol% 
THF -30°C 
87 3 
Fe(acac)3 1mol% 
THF -30°C 
75* 4 
FeCl3 3mol% 
THF -30°C 
83* 5 
NiCl2(dpp p) 1mol% 
THF -30°C 
72* 6 
CuCl3 100mol% 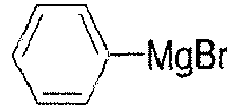
THF -30°C 
64* 7 
CoCl2 1mol% 
THF -30°C 
23* 8 
Li2CuCl4 5mol% 
THF -30°C 
68* 9 
Fe(acac)3 3mol% 
THF -30°C 
82* 10 
Fe(acac)3 3mol% 
THF -30°C 
83* * Determined by 19F NMR Table 2 Example Compound (1) Catalyst Compound (2) Solvent Temp. °C Product Yield (%)* 11 
Fe(acac)3 3mol% 
THF -30°C 
85* 12 
Fe(acac)3 3mol% 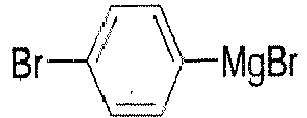
THF -30°C 
78* 13 
Fe(acac)3 3mol% 
THF -30°C 
80* 14 
Fe(acac)3 3mol% 
THF -30°C 
77* 15 
Fe(acac)3 3mol% 
THF -30°C 
75* 16 
Fe(acac)3 3mol% 
THF -30°C 
83 17 
Fe(acac)3 3mol% 
THF -30°C 
82 18 
no cat. 
THF -30°C 
20 19 
Fe(acac)3 3mol% 
THF -30°C 
45 *Determined by 19F NMR - Magnesium (1.6 g, 64.0 mmol), dibromoethane (0.6 g, 3.2 mmol), and cyclopentyl methyl ether (CPME) (10.0 mL) were added in a nitrogen atmosphere to a 50 ml three-necked flask equipped with a stirring bar and a thermometer. After bromocyclohexane (10.4 g, 64 mmol) was added, the mixture was stirred at 60°C for 1 hour to produce a Grignard compound.
- Subsequently, phenyl chlorothionoformate (11.0 g, 64 mmol), a 0.1 M tetrahydrofuran solution of dilithium tetrachlorocuprate(II)(19 mL, 1.9 mmol), and THF (10 mL) were added in a nitrogen atmosphere to a 200 ml three-necked flask equipped with a stirring bar and a thermometer. After the mixture was cooled to -30°C, the Grignard compound obtained by the above process was added, and the resulting mixture was stirred for 2 hours to complete the reaction. This mixture was returned to room temperature, and extracted with toluene/hydrochloric acid. The organic layer was evaporated under reduced pressure. The obtained solid was purified by silica gel column chromatography to give cyclohexyl thionobenzoate as a yellow solid (10.1 g, 46 mmol). The yield was 72%.
- Using the starting materials shown in Table 3, the reaction was performed in the same manner as in Example 20. Table 3 shows the kinds and yields of the obtained products. The each Grignard compound shown in the Compound (2) column of the table was obtained by reacting magnesium with a halide serving as precursor of each Grignard compound in the same manner as in Example 20. Further, the numerical values shown in the Catalyst column of the table are the amounts of catalyst expressed in mol% relative to Compound (1).
Table 3 Example Compound (1) Catalyst Compound (2) Solvent Temp. °C Product Yield (%)* 20 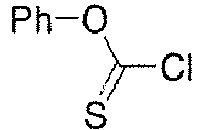
Li2CuCl4 3mol% 
CPME/THF -30°C 
72 21 
Li2CuCl4 3mol% 
CPME/THF -30°C 
65 22 
Li2CuCl4 3mol% 
CPME/THF -30°C 
68* 23 
Li2CuCl4 3mol% 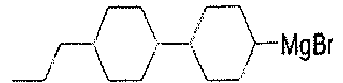
CPME/THF -30°C 
62 * Determined by 19F NMR
Claims (5)
- A process for producing an arylthionocarboxylic acid ester represented by Formula (3):

the process comprising reacting a halogenated thiocarbonyl compound represented by Formula (1):
- The process according to claim 1, wherein the reaction is performed in an aprotic organic solvent.
- The process according to claim 1 or 2, wherein the halogenated thiocarbonyl compound represented by Formula (1) is reacted with the Grignard compound represented by Formula (2) in the presence of at least one component selected from the group consisting of trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, and divalent cobalt compounds.
- A process for producing an arylthionocarboxylic acid ester represented by Formula (3):

the process comprising reacting a halide represented by Formula (4): R-Y wherein R is as defined above and Y is a halogen atom, with a magnesium compound, and then reacting the reaction product with a halogenated thiocarbonyl compound represented by Formula (1):
- The process according to claim 4, wherein the reaction with the halogenated thiocarbonyl compound represented by Formula (1) is performed in the presence of at least one component selected from the group consisting of trivalent iron compounds, monovalent copper compounds, divalent copper compounds, divalent nickel compounds, divalent zinc compounds, and divalent cobalt compounds.
Applications Claiming Priority (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2012213968A JP5737255B2 (en) | 2012-09-27 | 2012-09-27 | Method for producing thionocarboxylic acid ester |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| EP2712858A1 true EP2712858A1 (en) | 2014-04-02 |
| EP2712858B1 EP2712858B1 (en) | 2018-03-07 |
Family
ID=49165625
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP13184535.6A Active EP2712858B1 (en) | 2012-09-27 | 2013-09-16 | Process for producing thionocarboxylic acid ester |
Country Status (4)
| Country | Link |
|---|---|
| EP (1) | EP2712858B1 (en) |
| JP (1) | JP5737255B2 (en) |
| KR (1) | KR101546144B1 (en) |
| CN (1) | CN103694154B (en) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US10207976B2 (en) | 2015-09-30 | 2019-02-19 | Taisho Pharmaceutical Co., Ltd | Method for producing optically active 2-(2-fluorobiphenyl-4-yl) propanoic acid |
Families Citing this family (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2015141811A1 (en) * | 2014-03-20 | 2015-09-24 | ダイキン工業株式会社 | Method for producing compound having oxydifluoromethylene skeleton |
| EP3196183B1 (en) * | 2014-09-19 | 2019-05-01 | Toray Fine Chemicals Co., Ltd. | Method for producing 2'-trifluoromethyl group-substituted aromatic ketone |
Family Cites Families (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| MY113806A (en) * | 1995-07-21 | 2002-05-31 | Upjohn Co | Antiparasitic marcfortines and paraherquamides |
| CN101775102B (en) * | 2010-01-29 | 2012-06-06 | 湘潭大学 | Method for fast eliminating disulfide ester end groups of RAFT polymers |
-
2012
- 2012-09-27 JP JP2012213968A patent/JP5737255B2/en active Active
-
2013
- 2013-09-16 EP EP13184535.6A patent/EP2712858B1/en active Active
- 2013-09-24 KR KR1020130113304A patent/KR101546144B1/en active Active
- 2013-09-27 CN CN201310449678.3A patent/CN103694154B/en active Active
Non-Patent Citations (13)
| Title |
|---|
| A. OHNO ET AL., TETRAHEDRON LETT., 1968, pages 2083 |
| BULL. CHEM. SOC. BELG., vol. 87, 1987, pages 293 |
| C. CARDELLICCHIO ET AL: "A highly efficient synthetic route to ketones through sequential coupling reactions of grignard reagents with -phenyl carbonochloridothioate in the presence of nickel or iron catalysts", TETRAHEDRON LETTERS, vol. 26, no. 30, 1 January 1985 (1985-01-01), pages 3595 - 3598, XP055084505, ISSN: 0040-4039, DOI: 10.1016/S0040-4039(00)89199-6 * |
| H. VIOLA ET AL., CHEM. BER., vol. 101, 1968, pages 3517 |
| HORST VIOLA ET AL: "Organische Schwefelverbindungen,97. Friedel-Crafts-Reaktionen mit Thiosäurechloriden", CHEMISCHE BERICHTE, vol. 101, no. 10, 1 October 1968 (1968-10-01), pages 3517 - 3529, XP055084572, ISSN: 0009-2940, DOI: 10.1002/cber.19681011024 * |
| J. R. FALCK ET AL: "Tin-Copper Transmetalation: Cross-Coupling of .alpha.-Heteroatom-Substituted Alkyltributylstannanes with Organohalides", JOURNAL OF THE AMERICAN CHEMICAL SOCIETY, vol. 117, no. 22, 1 June 1995 (1995-06-01), pages 5973 - 5982, XP055084573, ISSN: 0002-7863, DOI: 10.1021/ja00127a010 * |
| K.A. LATIF ET AL., TETRAHEDRON, vol. 26, 1970, pages 4247 |
| KLAUS HARTKE ET AL: "[alpha],[beta]-acetylenic dithio and thiono esters", TETRAHEDRON LETTERS, vol. 30, no. 9, 1 January 1989 (1989-01-01), pages 1073 - 1076, XP055084574, ISSN: 0040-4039, DOI: 10.1016/S0040-4039(01)80363-4 * |
| LIEBIGS ANN. CHEM., 1974, pages 671 |
| LIEBIGS, ANN. CHEM., 1973, pages 1637 |
| S. SCHEITHAUER ET AL., CHEM. BER., vol. 98, 1965, pages 838 |
| SYNTHESIS, 1973, pages 149 |
| Z. CHEM., vol. 6, 1966, pages 108 |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US10207976B2 (en) | 2015-09-30 | 2019-02-19 | Taisho Pharmaceutical Co., Ltd | Method for producing optically active 2-(2-fluorobiphenyl-4-yl) propanoic acid |
Also Published As
| Publication number | Publication date |
|---|---|
| CN103694154B (en) | 2016-05-11 |
| JP2014065693A (en) | 2014-04-17 |
| KR20140041353A (en) | 2014-04-04 |
| CN103694154A (en) | 2014-04-02 |
| JP5737255B2 (en) | 2015-06-17 |
| EP2712858B1 (en) | 2018-03-07 |
| KR101546144B1 (en) | 2015-08-20 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| US10421702B2 (en) | Method for producing fluorine-containing olefin | |
| JP2012504153A5 (en) | ||
| CN101878192B (en) | Method for producing 4-perfluoroisopropylaniline | |
| EP2712858B1 (en) | Process for producing thionocarboxylic acid ester | |
| EP2711358B1 (en) | Process for producing aryl ester of thionocarboxylic acid | |
| Seyferth et al. | High-yield acyl anion trapping reactions. Nucleophilic acylation of dialkyl disulfides, S-alkyl thioesters and elemental sulfur | |
| EP3121160A1 (en) | Method for producing compound having oxydifluoromethylene skeleton | |
| EP0590539B1 (en) | Processes for producing tetrathiafulvalene derivatives and their precursors | |
| US20230130333A1 (en) | Method for manufacturing phosphonate esters and method for manufacturing phosphate esters | |
| JP5294303B2 (en) | Novel N, N, P-tridentate Schiff base ligand compound and asymmetric synthesis using the compound | |
| JP2008239584A (en) | Process for producing organophosphine complex of zerovalent nickel | |
| JPS62258333A (en) | Manufacture of aromatic aldehyde | |
| JP5488786B2 (en) | Process for producing azaboracyclopentene compound and synthetic intermediate thereof | |
| JP2014005214A (en) | Aryldichlorophosphine production method | |
| EP3239125A1 (en) | Method for producing olefin | |
| US11891409B2 (en) | Trifluoromethyl alkenylphosphonate and preparation method therefor | |
| CN115260103B (en) | Preparation method of 4,5-dihalogen-1- (difluoromethyl) -1H-imidazole | |
| KR100884636B1 (en) | Manufacturing method of coenzyme # 10 | |
| KR100855310B1 (en) | Method for preparing para-hydroquinone derivative | |
| KR20240060025A (en) | Manufacturing method of n-aryl amide compound | |
| KR20240073969A (en) | Method for producing organo-titanium compounds | |
| EP1883470B1 (en) | Catalyst precursor for the production of olefins with an odd number of carbon atoms, process for its preparation and production method for such olefins | |
| KR20230060027A (en) | Method for preparing nonsymmetric phosphate based compound | |
| JP2006182739A (en) | Organic bismuth compound and process for producing the same | |
| JPH0674250B2 (en) | Method for producing thiocarbamate derivative |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| 17P | Request for examination filed |
Effective date: 20130916 |
|
| AK | Designated contracting states |
Kind code of ref document: A1 Designated state(s): AL AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HR HU IE IS IT LI LT LU LV MC MK MT NL NO PL PT RO RS SE SI SK SM TR |
|
| AX | Request for extension of the european patent |
Extension state: BA ME |
|
| RAP1 | Party data changed (applicant data changed or rights of an application transferred) |
Owner name: DAIKIN INDUSTRIES, LTD. |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: EXAMINATION IS IN PROGRESS |
|
| 17Q | First examination report despatched |
Effective date: 20170405 |
|
| GRAP | Despatch of communication of intention to grant a patent |
Free format text: ORIGINAL CODE: EPIDOSNIGR1 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: GRANT OF PATENT IS INTENDED |
|
| INTG | Intention to grant announced |
Effective date: 20170913 |
|
| GRAJ | Information related to disapproval of communication of intention to grant by the applicant or resumption of examination proceedings by the epo deleted |
Free format text: ORIGINAL CODE: EPIDOSDIGR1 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: EXAMINATION IS IN PROGRESS |
|
| GRAR | Information related to intention to grant a patent recorded |
Free format text: ORIGINAL CODE: EPIDOSNIGR71 |
|
| GRAS | Grant fee paid |
Free format text: ORIGINAL CODE: EPIDOSNIGR3 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: GRANT OF PATENT IS INTENDED |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: THE PATENT HAS BEEN GRANTED |
|
| INTC | Intention to grant announced (deleted) | ||
| AK | Designated contracting states |
Kind code of ref document: B1 Designated state(s): AL AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HR HU IE IS IT LI LT LU LV MC MK MT NL NO PL PT RO RS SE SI SK SM TR |
|
| INTG | Intention to grant announced |
Effective date: 20180126 |
|
| REG | Reference to a national code |
Ref country code: GB Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: EP Ref country code: AT Ref legal event code: REF Ref document number: 976400 Country of ref document: AT Kind code of ref document: T Effective date: 20180315 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R096 Ref document number: 602013033959 Country of ref document: DE |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: NL Ref legal event code: MP Effective date: 20180307 |
|
| REG | Reference to a national code |
Ref country code: LT Ref legal event code: MG4D |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: CY Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: LT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: HR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: NO Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180607 Ref country code: ES Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: FI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| REG | Reference to a national code |
Ref country code: FR Ref legal event code: PLFP Year of fee payment: 6 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: RS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: SE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: LV Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: GR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180608 Ref country code: BG Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180607 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: PL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: AL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: EE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: RO Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: NL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: SM Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 Ref country code: CZ Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| REG | Reference to a national code |
Ref country code: DE Ref legal event code: R097 Ref document number: 602013033959 Country of ref document: DE |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: PT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180709 |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| 26N | No opposition filed |
Effective date: 20181210 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MC Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| REG | Reference to a national code |
Ref country code: BE Ref legal event code: MM Effective date: 20180930 |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: MM4A |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LU Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20180916 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20180916 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: BE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20180930 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MT Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20180916 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: TR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180307 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: HU Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT; INVALID AB INITIO Effective date: 20130916 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MK Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20180307 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IS Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20180707 |
|
| REG | Reference to a national code |
Ref country code: AT Ref legal event code: UEP Ref document number: 976400 Country of ref document: AT Kind code of ref document: T Effective date: 20180307 |
|
| P01 | Opt-out of the competence of the unified patent court (upc) registered |
Effective date: 20230525 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: U11 Free format text: ST27 STATUS EVENT CODE: U-0-0-U10-U11 (AS PROVIDED BY THE NATIONAL OFFICE) Effective date: 20251001 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 20250730 Year of fee payment: 13 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: IT Payment date: 20250825 Year of fee payment: 13 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: GB Payment date: 20250731 Year of fee payment: 13 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: FR Payment date: 20250808 Year of fee payment: 13 Ref country code: AT Payment date: 20250827 Year of fee payment: 13 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: CH Payment date: 20251001 Year of fee payment: 13 |