EP0894093B1 - Soluble tachykinin antagonists, the preparation and use thereof - Google Patents
Soluble tachykinin antagonists, the preparation and use thereof Download PDFInfo
- Publication number
- EP0894093B1 EP0894093B1 EP97905089A EP97905089A EP0894093B1 EP 0894093 B1 EP0894093 B1 EP 0894093B1 EP 97905089 A EP97905089 A EP 97905089A EP 97905089 A EP97905089 A EP 97905089A EP 0894093 B1 EP0894093 B1 EP 0894093B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- asp
- dap
- compounds
- cyclo
- cysteine
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Expired - Lifetime
Links
- 239000005557 antagonist Substances 0.000 title description 11
- 238000002360 preparation method Methods 0.000 title description 8
- 108060008037 tachykinin Proteins 0.000 title description 7
- 102000003141 Tachykinin Human genes 0.000 title description 6
- 150000001875 compounds Chemical class 0.000 claims abstract description 31
- 235000001014 amino acid Nutrition 0.000 claims abstract description 13
- 150000001413 amino acids Chemical class 0.000 claims abstract description 12
- 235000018417 cysteine Nutrition 0.000 claims abstract description 8
- XUJNEKJLAYXESH-UHFFFAOYSA-N cysteine Natural products SCC(N)C(O)=O XUJNEKJLAYXESH-UHFFFAOYSA-N 0.000 claims abstract description 8
- 230000002209 hydrophobic effect Effects 0.000 claims abstract description 5
- 239000004372 Polyvinyl alcohol Substances 0.000 claims abstract description 3
- 150000002243 furanoses Chemical class 0.000 claims abstract description 3
- LNEPOXFFQSENCJ-UHFFFAOYSA-N haloperidol Chemical compound C1CC(O)(C=2C=CC(Cl)=CC=2)CCN1CCCC(=O)C1=CC=C(F)C=C1 LNEPOXFFQSENCJ-UHFFFAOYSA-N 0.000 claims abstract description 3
- 150000002574 ketohexoses Chemical class 0.000 claims abstract description 3
- 229920002451 polyvinyl alcohol Polymers 0.000 claims abstract description 3
- 125000003130 L-selenocysteinyl group Chemical group O=C([*])[C@@](N([H])[H])([H])C([H])([H])[Se][H] 0.000 claims abstract 3
- JULROCUWKLNBSN-UHFFFAOYSA-N seleno-DL-cystine Natural products OC(=O)C(N)C[Se][Se]CC(N)C(O)=O JULROCUWKLNBSN-UHFFFAOYSA-N 0.000 claims abstract 3
- 150000003215 pyranoses Chemical group 0.000 claims abstract 2
- 239000008194 pharmaceutical composition Substances 0.000 claims description 5
- 239000003814 drug Substances 0.000 claims description 3
- 206010007270 Carcinoid syndrome Diseases 0.000 claims description 2
- 206010052402 Gastrointestinal hypermotility Diseases 0.000 claims description 2
- 208000023105 Huntington disease Diseases 0.000 claims description 2
- 206010020772 Hypertension Diseases 0.000 claims description 2
- 206010061218 Inflammation Diseases 0.000 claims description 2
- 208000019695 Migraine disease Diseases 0.000 claims description 2
- 206010029240 Neuritis Diseases 0.000 claims description 2
- 206010046543 Urinary incontinence Diseases 0.000 claims description 2
- 208000024780 Urticaria Diseases 0.000 claims description 2
- 206010003246 arthritis Diseases 0.000 claims description 2
- 208000006673 asthma Diseases 0.000 claims description 2
- 230000004054 inflammatory process Effects 0.000 claims description 2
- 206010022000 influenza Diseases 0.000 claims description 2
- 206010027599 migraine Diseases 0.000 claims description 2
- 208000004296 neuralgia Diseases 0.000 claims description 2
- 208000024891 symptom Diseases 0.000 claims description 2
- 230000001173 tumoral effect Effects 0.000 claims description 2
- 230000000694 effects Effects 0.000 abstract description 7
- 230000003389 potentiating effect Effects 0.000 abstract description 2
- 101000713585 Homo sapiens Tubulin beta-4A chain Proteins 0.000 abstract 1
- 102100036788 Tubulin beta-4A chain Human genes 0.000 abstract 1
- 108090000765 processed proteins & peptides Proteins 0.000 description 24
- WQZGKKKJIJFFOK-FPRJBGLDSA-N beta-D-galactose Chemical compound OC[C@H]1O[C@@H](O)[C@H](O)[C@@H](O)[C@H]1O WQZGKKKJIJFFOK-FPRJBGLDSA-N 0.000 description 22
- PECYZEOJVXMISF-UHFFFAOYSA-N 3-aminoalanine Chemical compound [NH3+]CC(N)C([O-])=O PECYZEOJVXMISF-UHFFFAOYSA-N 0.000 description 20
- 239000000243 solution Substances 0.000 description 17
- JGFZNNIVVJXRND-UHFFFAOYSA-N N,N-Diisopropylethylamine (DIPEA) Chemical compound CCN(C(C)C)C(C)C JGFZNNIVVJXRND-UHFFFAOYSA-N 0.000 description 16
- YMWUJEATGCHHMB-UHFFFAOYSA-N Dichloromethane Chemical compound ClCCl YMWUJEATGCHHMB-UHFFFAOYSA-N 0.000 description 15
- OKKJLVBELUTLKV-UHFFFAOYSA-N Methanol Chemical compound OC OKKJLVBELUTLKV-UHFFFAOYSA-N 0.000 description 15
- ZMXDDKWLCZADIW-UHFFFAOYSA-N N,N-Dimethylformamide Chemical compound CN(C)C=O ZMXDDKWLCZADIW-UHFFFAOYSA-N 0.000 description 13
- XLYOFNOQVPJJNP-UHFFFAOYSA-N water Substances O XLYOFNOQVPJJNP-UHFFFAOYSA-N 0.000 description 12
- 238000000034 method Methods 0.000 description 11
- 239000011347 resin Substances 0.000 description 10
- 229920005989 resin Polymers 0.000 description 10
- 239000000047 product Substances 0.000 description 9
- DTQVDTLACAAQTR-UHFFFAOYSA-N Trifluoroacetic acid Chemical compound OC(=O)C(F)(F)F DTQVDTLACAAQTR-UHFFFAOYSA-N 0.000 description 8
- 238000010511 deprotection reaction Methods 0.000 description 8
- FDKWRPBBCBCIGA-REOHCLBHSA-N (2r)-2-azaniumyl-3-$l^{1}-selanylpropanoate Chemical compound [Se]C[C@H](N)C(O)=O FDKWRPBBCBCIGA-REOHCLBHSA-N 0.000 description 7
- FDKWRPBBCBCIGA-UWTATZPHSA-N D-Selenocysteine Natural products [Se]C[C@@H](N)C(O)=O FDKWRPBBCBCIGA-UWTATZPHSA-N 0.000 description 7
- 238000006243 chemical reaction Methods 0.000 description 7
- 238000004128 high performance liquid chromatography Methods 0.000 description 7
- 238000007363 ring formation reaction Methods 0.000 description 7
- ZKZBPNGNEQAJSX-UHFFFAOYSA-N selenocysteine Natural products [SeH]CC(N)C(O)=O ZKZBPNGNEQAJSX-UHFFFAOYSA-N 0.000 description 7
- 235000016491 selenocysteine Nutrition 0.000 description 7
- 229940055619 selenocysteine Drugs 0.000 description 7
- QTBSBXVTEAMEQO-UHFFFAOYSA-N Acetic acid Chemical compound CC(O)=O QTBSBXVTEAMEQO-UHFFFAOYSA-N 0.000 description 6
- KFZMGEQAYNKOFK-UHFFFAOYSA-N Isopropanol Chemical compound CC(C)O KFZMGEQAYNKOFK-UHFFFAOYSA-N 0.000 description 6
- -1 MEN 10627 compound Chemical class 0.000 description 6
- 238000002953 preparative HPLC Methods 0.000 description 6
- QOSSAOTZNIDXMA-UHFFFAOYSA-N Dicylcohexylcarbodiimide Chemical compound C1CCCCC1N=C=NC1CCCCC1 QOSSAOTZNIDXMA-UHFFFAOYSA-N 0.000 description 5
- TUCNEACPLKLKNU-UHFFFAOYSA-N acetyl Chemical compound C[C]=O TUCNEACPLKLKNU-UHFFFAOYSA-N 0.000 description 5
- 230000004071 biological effect Effects 0.000 description 5
- 230000008878 coupling Effects 0.000 description 5
- 238000010168 coupling process Methods 0.000 description 5
- 238000005859 coupling reaction Methods 0.000 description 5
- 238000004992 fast atom bombardment mass spectroscopy Methods 0.000 description 5
- 102000004196 processed proteins & peptides Human genes 0.000 description 5
- 239000007790 solid phase Substances 0.000 description 5
- ASOKPJOREAFHNY-UHFFFAOYSA-N 1-Hydroxybenzotriazole Chemical compound C1=CC=C2N(O)N=NC2=C1 ASOKPJOREAFHNY-UHFFFAOYSA-N 0.000 description 4
- 239000002202 Polyethylene glycol Substances 0.000 description 4
- 230000000052 comparative effect Effects 0.000 description 4
- 239000012043 crude product Substances 0.000 description 4
- NPZTUJOABDZTLV-UHFFFAOYSA-N hydroxybenzotriazole Substances O=C1C=CC=C2NNN=C12 NPZTUJOABDZTLV-UHFFFAOYSA-N 0.000 description 4
- 125000002950 monocyclic group Chemical group 0.000 description 4
- IWDCLRJOBJJRNH-UHFFFAOYSA-N p-cresol Chemical compound CC1=CC=C(O)C=C1 IWDCLRJOBJJRNH-UHFFFAOYSA-N 0.000 description 4
- 230000000144 pharmacologic effect Effects 0.000 description 4
- 229920001223 polyethylene glycol Polymers 0.000 description 4
- 125000005931 tert-butyloxycarbonyl group Chemical group [H]C([H])([H])C(OC(*)=O)(C([H])([H])[H])C([H])([H])[H] 0.000 description 4
- UKAUYVFTDYCKQA-UHFFFAOYSA-N -2-Amino-4-hydroxybutanoic acid Natural products OC(=O)C(N)CCO UKAUYVFTDYCKQA-UHFFFAOYSA-N 0.000 description 3
- FFFHZYDWPBMWHY-VKHMYHEASA-N L-homocysteine Chemical compound OC(=O)[C@@H](N)CCS FFFHZYDWPBMWHY-VKHMYHEASA-N 0.000 description 3
- UKAUYVFTDYCKQA-VKHMYHEASA-N L-homoserine Chemical compound OC(=O)[C@@H](N)CCO UKAUYVFTDYCKQA-VKHMYHEASA-N 0.000 description 3
- 108010072184 MEN 10627 Proteins 0.000 description 3
- HEMHJVSKTPXQMS-UHFFFAOYSA-M Sodium hydroxide Chemical compound [OH-].[Na+] HEMHJVSKTPXQMS-UHFFFAOYSA-M 0.000 description 3
- 102100037342 Substance-K receptor Human genes 0.000 description 3
- 238000004458 analytical method Methods 0.000 description 3
- 210000004899 c-terminal region Anatomy 0.000 description 3
- 150000001793 charged compounds Chemical class 0.000 description 3
- 230000000875 corresponding effect Effects 0.000 description 3
- 239000007822 coupling agent Substances 0.000 description 3
- 229930182830 galactose Natural products 0.000 description 3
- 238000000746 purification Methods 0.000 description 3
- 230000004044 response Effects 0.000 description 3
- QDZOEBFLNHCSSF-PFFBOGFISA-N (2S)-2-[[(2R)-2-[[(2S)-1-[(2S)-6-amino-2-[[(2S)-1-[(2R)-2-amino-5-carbamimidamidopentanoyl]pyrrolidine-2-carbonyl]amino]hexanoyl]pyrrolidine-2-carbonyl]amino]-3-(1H-indol-3-yl)propanoyl]amino]-N-[(2R)-1-[[(2S)-1-[[(2R)-1-[[(2S)-1-[[(2S)-1-amino-4-methyl-1-oxopentan-2-yl]amino]-4-methyl-1-oxopentan-2-yl]amino]-3-(1H-indol-3-yl)-1-oxopropan-2-yl]amino]-1-oxo-3-phenylpropan-2-yl]amino]-3-(1H-indol-3-yl)-1-oxopropan-2-yl]pentanediamide Chemical compound C([C@@H](C(=O)N[C@H](CC=1C2=CC=CC=C2NC=1)C(=O)N[C@@H](CC(C)C)C(=O)N[C@@H](CC(C)C)C(N)=O)NC(=O)[C@@H](CC=1C2=CC=CC=C2NC=1)NC(=O)[C@H](CCC(N)=O)NC(=O)[C@@H](CC=1C2=CC=CC=C2NC=1)NC(=O)[C@H]1N(CCC1)C(=O)[C@H](CCCCN)NC(=O)[C@H]1N(CCC1)C(=O)[C@H](N)CCCNC(N)=N)C1=CC=CC=C1 QDZOEBFLNHCSSF-PFFBOGFISA-N 0.000 description 2
- HEDRZPFGACZZDS-UHFFFAOYSA-N Chloroform Chemical compound ClC(Cl)Cl HEDRZPFGACZZDS-UHFFFAOYSA-N 0.000 description 2
- KRHYYFGTRYWZRS-UHFFFAOYSA-N Fluorane Chemical compound F KRHYYFGTRYWZRS-UHFFFAOYSA-N 0.000 description 2
- HEAUFJZALFKPBA-YRVBCFNBSA-N Neurokinin A Chemical compound C([C@@H](C(=O)N[C@H](C(=O)NCC(=O)N[C@@H](CC(C)C)C(=O)N[C@@H](CCSC)C(N)=O)C(C)C)NC(=O)[C@H](CO)NC(=O)[C@H](CC(O)=O)NC(=O)[C@@H](NC(=O)[C@H](CCCCN)NC(=O)[C@@H](N)CC=1NC=NC=1)C(C)O)C1=CC=CC=C1 HEAUFJZALFKPBA-YRVBCFNBSA-N 0.000 description 2
- 102400000097 Neurokinin A Human genes 0.000 description 2
- 101800000399 Neurokinin A Proteins 0.000 description 2
- NQRYJNQNLNOLGT-UHFFFAOYSA-N Piperidine Chemical compound C1CCNCC1 NQRYJNQNLNOLGT-UHFFFAOYSA-N 0.000 description 2
- 102400000096 Substance P Human genes 0.000 description 2
- 101800003906 Substance P Proteins 0.000 description 2
- 101710116609 Substance-K receptor Proteins 0.000 description 2
- 239000000556 agonist Substances 0.000 description 2
- 230000015572 biosynthetic process Effects 0.000 description 2
- 239000000706 filtrate Substances 0.000 description 2
- 239000012634 fragment Substances 0.000 description 2
- 125000000524 functional group Chemical group 0.000 description 2
- 238000000338 in vitro Methods 0.000 description 2
- 239000000203 mixture Substances 0.000 description 2
- 238000010647 peptide synthesis reaction Methods 0.000 description 2
- 125000002924 primary amino group Chemical group [H]N([H])* 0.000 description 2
- 238000010532 solid phase synthesis reaction Methods 0.000 description 2
- 239000002904 solvent Substances 0.000 description 2
- 238000003786 synthesis reaction Methods 0.000 description 2
- 238000010189 synthetic method Methods 0.000 description 2
- 150000003568 thioethers Chemical class 0.000 description 2
- 238000005406 washing Methods 0.000 description 2
- WQZGKKKJIJFFOK-SVZMEOIVSA-N (+)-Galactose Chemical compound OC[C@H]1OC(O)[C@H](O)[C@@H](O)[C@H]1O WQZGKKKJIJFFOK-SVZMEOIVSA-N 0.000 description 1
- CFDNUNSOUUFTQO-JYMVZIKVSA-N (1r,4s,7s,10r,13s,16r)-13-benzyl-16-(1h-indol-3-ylmethyl)-7-(2-methylpropyl)-4-(2-methylsulfanylethyl)-2,5,8,11,14,17,21-heptazabicyclo[8.8.4]docosane-3,6,9,12,15,18,20-heptone Chemical compound C([C@@H]1NC(=O)[C@@H](CC=2C3=CC=CC=C3NC=2)NC(=O)[C@H]2CC(=O)NC[C@@H](NC1=O)C(=O)N[C@@H](CC(C)C)C(=O)N[C@H](C(N2)=O)CCSC)C1=CC=CC=C1 CFDNUNSOUUFTQO-JYMVZIKVSA-N 0.000 description 1
- MGHMWKZOLAAOTD-DEOSSOPVSA-N (2s)-2-(9h-fluoren-9-ylmethoxycarbonylamino)-3-(1h-indol-3-yl)propanoic acid Chemical compound C12=CC=CC=C2C2=CC=CC=C2C1COC(=O)N[C@H](C(=O)O)CC1=CNC2=CC=CC=C12 MGHMWKZOLAAOTD-DEOSSOPVSA-N 0.000 description 1
- PKAUMAVONPSDRW-IBGZPJMESA-N (2s)-2-(9h-fluoren-9-ylmethoxycarbonylamino)-3-[(2-methylpropan-2-yl)oxycarbonylamino]propanoic acid Chemical compound C1=CC=C2C(COC(=O)N[C@@H](CNC(=O)OC(C)(C)C)C(O)=O)C3=CC=CC=C3C2=C1 PKAUMAVONPSDRW-IBGZPJMESA-N 0.000 description 1
- SJVFAHZPLIXNDH-QFIPXVFZSA-N (2s)-2-(9h-fluoren-9-ylmethoxycarbonylamino)-3-phenylpropanoic acid Chemical compound C([C@@H](C(=O)O)NC(=O)OCC1C2=CC=CC=C2C2=CC=CC=C21)C1=CC=CC=C1 SJVFAHZPLIXNDH-QFIPXVFZSA-N 0.000 description 1
- FODJWPHPWBKDON-IBGZPJMESA-N (2s)-2-(9h-fluoren-9-ylmethoxycarbonylamino)-4-[(2-methylpropan-2-yl)oxy]-4-oxobutanoic acid Chemical compound C1=CC=C2C(COC(=O)N[C@@H](CC(=O)OC(C)(C)C)C(O)=O)C3=CC=CC=C3C2=C1 FODJWPHPWBKDON-IBGZPJMESA-N 0.000 description 1
- BVAUMRCGVHUWOZ-ZETCQYMHSA-N (2s)-2-(cyclohexylazaniumyl)propanoate Chemical compound OC(=O)[C@H](C)NC1CCCCC1 BVAUMRCGVHUWOZ-ZETCQYMHSA-N 0.000 description 1
- IYKLZBIWFXPUCS-VIFPVBQESA-N (2s)-2-(naphthalen-1-ylamino)propanoic acid Chemical compound C1=CC=C2C(N[C@@H](C)C(O)=O)=CC=CC2=C1 IYKLZBIWFXPUCS-VIFPVBQESA-N 0.000 description 1
- ZYJPUMXJBDHSIF-NSHDSACASA-N (2s)-2-[(2-methylpropan-2-yl)oxycarbonylamino]-3-phenylpropanoic acid Chemical compound CC(C)(C)OC(=O)N[C@H](C(O)=O)CC1=CC=CC=C1 ZYJPUMXJBDHSIF-NSHDSACASA-N 0.000 description 1
- IHXHBYFWSOYYTR-ZDUSSCGKSA-N (2s)-3-(1-formylindol-3-yl)-2-[(2-methylpropan-2-yl)oxycarbonylamino]propanoic acid Chemical compound C1=CC=C2C(C[C@H](NC(=O)OC(C)(C)C)C(O)=O)=CN(C=O)C2=C1 IHXHBYFWSOYYTR-ZDUSSCGKSA-N 0.000 description 1
- MVWPBNQGEGBGRF-IBGZPJMESA-N (2s)-3-(9h-fluoren-9-ylmethoxycarbonylamino)-2-[(2-methylpropan-2-yl)oxycarbonylamino]propanoic acid Chemical compound C1=CC=C2C(COC(=O)NC[C@H](NC(=O)OC(C)(C)C)C(O)=O)C3=CC=CC=C3C2=C1 MVWPBNQGEGBGRF-IBGZPJMESA-N 0.000 description 1
- NHLRMCFWGFPSLT-IBGZPJMESA-N (2s)-4-(9h-fluoren-9-ylmethoxy)-2-[(2-methylpropan-2-yl)oxycarbonylamino]-4-oxobutanoic acid Chemical compound C1=CC=C2C(COC(=O)C[C@H](NC(=O)OC(C)(C)C)C(O)=O)C3=CC=CC=C3C2=C1 NHLRMCFWGFPSLT-IBGZPJMESA-N 0.000 description 1
- YEDUAINPPJYDJZ-UHFFFAOYSA-N 2-hydroxybenzothiazole Chemical compound C1=CC=C2SC(O)=NC2=C1 YEDUAINPPJYDJZ-UHFFFAOYSA-N 0.000 description 1
- WWYFPDXEIFBNKE-UHFFFAOYSA-N 4-(hydroxymethyl)benzoic acid Chemical compound OCC1=CC=C(C(O)=O)C=C1 WWYFPDXEIFBNKE-UHFFFAOYSA-N 0.000 description 1
- WLHCBQAPPJAULW-UHFFFAOYSA-N 4-methylbenzenethiol Chemical compound CC1=CC=C(S)C=C1 WLHCBQAPPJAULW-UHFFFAOYSA-N 0.000 description 1
- HBAQYPYDRFILMT-UHFFFAOYSA-N 8-[3-(1-cyclopropylpyrazol-4-yl)-1H-pyrazolo[4,3-d]pyrimidin-5-yl]-3-methyl-3,8-diazabicyclo[3.2.1]octan-2-one Chemical class C1(CC1)N1N=CC(=C1)C1=NNC2=C1N=C(N=C2)N1C2C(N(CC1CC2)C)=O HBAQYPYDRFILMT-UHFFFAOYSA-N 0.000 description 1
- WEVYAHXRMPXWCK-UHFFFAOYSA-N Acetonitrile Chemical compound CC#N WEVYAHXRMPXWCK-UHFFFAOYSA-N 0.000 description 1
- QNAYBMKLOCPYGJ-UHFFFAOYSA-N D-alpha-Ala Natural products CC([NH3+])C([O-])=O QNAYBMKLOCPYGJ-UHFFFAOYSA-N 0.000 description 1
- QMMFVYPAHWMCMS-UHFFFAOYSA-N Dimethyl sulfide Chemical compound CSC QMMFVYPAHWMCMS-UHFFFAOYSA-N 0.000 description 1
- WQZGKKKJIJFFOK-GASJEMHNSA-N Glucose Chemical group OC[C@H]1OC(O)[C@H](O)[C@@H](O)[C@@H]1O WQZGKKKJIJFFOK-GASJEMHNSA-N 0.000 description 1
- SQUHHTBVTRBESD-UHFFFAOYSA-N Hexa-Ac-myo-Inositol Chemical group CC(=O)OC1C(OC(C)=O)C(OC(C)=O)C(OC(C)=O)C(OC(C)=O)C1OC(C)=O SQUHHTBVTRBESD-UHFFFAOYSA-N 0.000 description 1
- 241001465754 Metazoa Species 0.000 description 1
- MDXGYYOJGPFFJL-QMMMGPOBSA-N N(alpha)-t-butoxycarbonyl-L-leucine Chemical compound CC(C)C[C@@H](C(O)=O)NC(=O)OC(C)(C)C MDXGYYOJGPFFJL-QMMMGPOBSA-N 0.000 description 1
- 239000007832 Na2SO4 Substances 0.000 description 1
- 108010040722 Neurokinin-2 Receptors Proteins 0.000 description 1
- PMZURENOXWZQFD-UHFFFAOYSA-L Sodium Sulfate Chemical compound [Na+].[Na+].[O-]S([O-])(=O)=O PMZURENOXWZQFD-UHFFFAOYSA-L 0.000 description 1
- UIIMBOGNXHQVGW-UHFFFAOYSA-M Sodium bicarbonate Chemical class [Na+].OC([O-])=O UIIMBOGNXHQVGW-UHFFFAOYSA-M 0.000 description 1
- WQDUMFSSJAZKTM-UHFFFAOYSA-N Sodium methoxide Chemical compound [Na+].[O-]C WQDUMFSSJAZKTM-UHFFFAOYSA-N 0.000 description 1
- 125000002777 acetyl group Chemical group [H]C([H])([H])C(*)=O 0.000 description 1
- 230000002378 acidificating effect Effects 0.000 description 1
- 230000003213 activating effect Effects 0.000 description 1
- 239000000443 aerosol Substances 0.000 description 1
- WQZGKKKJIJFFOK-PHYPRBDBSA-N alpha-D-galactose Chemical group OC[C@H]1O[C@H](O)[C@H](O)[C@@H](O)[C@H]1O WQZGKKKJIJFFOK-PHYPRBDBSA-N 0.000 description 1
- 239000007864 aqueous solution Substances 0.000 description 1
- WQZGKKKJIJFFOK-VFUOTHLCSA-N beta-D-glucose Chemical group OC[C@H]1O[C@@H](O)[C@H](O)[C@@H](O)[C@@H]1O WQZGKKKJIJFFOK-VFUOTHLCSA-N 0.000 description 1
- UCMIRNVEIXFBKS-UHFFFAOYSA-N beta-alanine Chemical compound NCCC(O)=O UCMIRNVEIXFBKS-UHFFFAOYSA-N 0.000 description 1
- 125000002619 bicyclic group Chemical group 0.000 description 1
- 230000008512 biological response Effects 0.000 description 1
- 239000002775 capsule Substances 0.000 description 1
- 239000003795 chemical substances by application Substances 0.000 description 1
- 230000002596 correlated effect Effects 0.000 description 1
- 239000013058 crude material Substances 0.000 description 1
- 230000007423 decrease Effects 0.000 description 1
- 231100000673 dose–response relationship Toxicity 0.000 description 1
- 230000032050 esterification Effects 0.000 description 1
- 238000005886 esterification reaction Methods 0.000 description 1
- 238000009472 formulation Methods 0.000 description 1
- 239000008103 glucose Chemical group 0.000 description 1
- 230000007062 hydrolysis Effects 0.000 description 1
- 238000006460 hydrolysis reaction Methods 0.000 description 1
- 230000005764 inhibitory process Effects 0.000 description 1
- CDAISMWEOUEBRE-GPIVLXJGSA-N inositol Chemical group O[C@H]1[C@H](O)[C@@H](O)[C@H](O)[C@H](O)[C@@H]1O CDAISMWEOUEBRE-GPIVLXJGSA-N 0.000 description 1
- 229960000367 inositol Drugs 0.000 description 1
- 239000000543 intermediate Substances 0.000 description 1
- 238000002955 isolation Methods 0.000 description 1
- 230000014759 maintenance of location Effects 0.000 description 1
- 238000004519 manufacturing process Methods 0.000 description 1
- 239000012074 organic phase Substances 0.000 description 1
- 239000000825 pharmaceutical preparation Substances 0.000 description 1
- 230000000750 progressive effect Effects 0.000 description 1
- 235000018102 proteins Nutrition 0.000 description 1
- 102000004169 proteins and genes Human genes 0.000 description 1
- 108090000623 proteins and genes Proteins 0.000 description 1
- 150000003214 pyranose derivatives Chemical group 0.000 description 1
- CDAISMWEOUEBRE-UHFFFAOYSA-N scyllo-inosotol Chemical group OC1C(O)C(O)C(O)C(O)C1O CDAISMWEOUEBRE-UHFFFAOYSA-N 0.000 description 1
- 229910052938 sodium sulfate Inorganic materials 0.000 description 1
- 150000005846 sugar alcohols Polymers 0.000 description 1
- 239000006188 syrup Substances 0.000 description 1
- 235000020357 syrup Nutrition 0.000 description 1
- 239000003826 tablet Substances 0.000 description 1
- 230000001225 therapeutic effect Effects 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K7/04—Linear peptides containing only normal peptide links
- C07K7/22—Tachykinins, e.g. Eledoisins, Substance P; Related peptides
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K7/04—Linear peptides containing only normal peptide links
- C07K7/06—Linear peptides containing only normal peptide links having 5 to 11 amino acids
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
- A61P1/14—Prodigestives, e.g. acids, enzymes, appetite stimulants, antidyspeptics, tonics, antiflatulents
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P11/00—Drugs for disorders of the respiratory system
- A61P11/06—Antiasthmatics
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P13/00—Drugs for disorders of the urinary system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P17/00—Drugs for dermatological disorders
- A61P17/04—Antipruritics
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P19/00—Drugs for skeletal disorders
- A61P19/02—Drugs for skeletal disorders for joint disorders, e.g. arthritis, arthrosis
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/06—Antimigraine agents
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/14—Drugs for disorders of the nervous system for treating abnormal movements, e.g. chorea, dyskinesia
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P29/00—Non-central analgesic, antipyretic or antiinflammatory agents, e.g. antirheumatic agents; Non-steroidal antiinflammatory drugs [NSAID]
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P35/00—Antineoplastic agents
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P9/00—Drugs for disorders of the cardiovascular system
- A61P9/12—Antihypertensives
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K7/00—Peptides having 5 to 20 amino acids in a fully defined sequence; Derivatives thereof
- C07K7/50—Cyclic peptides containing at least one abnormal peptide link
- C07K7/54—Cyclic peptides containing at least one abnormal peptide link with at least one abnormal peptide link in the ring
- C07K7/56—Cyclic peptides containing at least one abnormal peptide link with at least one abnormal peptide link in the ring the cyclisation not occurring through 2,4-diamino-butanoic acid
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07K—PEPTIDES
- C07K9/00—Peptides having up to 20 amino acids, containing saccharide radicals and having a fully defined sequence; Derivatives thereof
- C07K9/006—Peptides having up to 20 amino acids, containing saccharide radicals and having a fully defined sequence; Derivatives thereof the peptide sequence being part of a ring structure
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K38/00—Medicinal preparations containing peptides
Definitions
- the present invention refers to soluble tachykinin antagonists, to their preparation and to pharmaceutical compositions containing them.
- the present invention discloses tachykinin-antagonists which at the same time present a high water solubility and a high biologic activity.
- the compounds of formula (I) have many chiral centres, and therefore the invention comprises all the possible enantiomers.
- the compounds of formula (I) show unique structural characteristics, which differentiate them from the known products.
- the claimed compounds contrary to those reported up to now (A. T. McKnight et al., Br. J. Pharmacol. 1991, 41, 376; C. A. Maggi et al., J. Pharmacol. and Expt. Therapeutics, 1994, 271, 1489), contain highly hydrophilic functional groups, which give them an at least 100 times higher water solubility.
- the high water solubility and the strong pharmacological activity, higher than that of the insoluble compounds previously developed, make this new class of molecules ideal for the development of a wide set of pharmaceutical formulations.
- the compounds of the invention substantially differ from the insoluble derivatives previously studied in that the latter do not permit the development of pharmaceutical compositions which can be used therapeutically.
- the compounds of formula I characterized by a residue of cysteine or of selenocysteine, whose respective -SH or -SeH group is involved in a thio- or seleno-glycosidic or thio- or seleno-ether bond with a galactose, glucose, inositol or PEG residue or low molecular weight polyalcohols, eventhough with high water solubility, they have an activity even a little higher than the activity of the best antagonists of the insoluble type, such as the MEN 10627 compound (C. A. Maggi et al., J. Pharmacol. and Expt. Therapeutics, 1994, 271, 1489; G.
- the MEN 10627 compound C. A. Maggi et al., J. Pharmacol. and Expt. Therapeutics, 1994, 271, 1489; G.
- the compounds of formula (I), which are the object of the present invention, can be synthesized according to the different techniques known from literature, e.g. Schroeder et al. "The Peptides” vol. 1, Academic. Press., 1965; Bodanszky et al. "Peptide Synthesis Interscience Public Shescher, 1966; Barany & Merrifield, "The peptides; Analysis, Synthesis, Biology", 2, Chapter 1, Academic. Press., 1980. These techniques include the peptide-solid phase synthesis, the peptide synthesis in solution, the synthetic methods of the organic chemistry, or any combination thereof. The chosen synthetic scheme will obviously depend on the composition of the particular molecule. Since all of the claimed molecules are peptides, synthetic methods based on appropriate combinations of solid phase techniques and the classic methods in solution are preferably used, which involve low costs of production particularly in an industrial scale. In details such methods consist in:
- Fmoc-Cys( ⁇ -D-Gal(Ac)4)-OH, Fmoc-Ser( ⁇ -D-Gal(Ac)4)-OH, Fmoc-Asn( ⁇ -D-Gal)-OH can be obtained according to the methods known in literature (M. Gerz, et al., Angew. Chem. Int. Ed. Engl., 1993, 32, 269; Kessler, H. et al. J. Am. Chem. Soc. 1992, 114, 4805; Kessler H. et al., J. Am. Chem. Soc. 1991, 113, 7550).
- the seleno-derivatives are obtained with similar procedures starting from the selenocysteine, using the known techniques for the protection of the amino function.
- these compounds are suitable for the therapeutic administration to the superior animals and to humans.
- their water solubility allows to prepare simple and unexpensive pharmaceutical preparations, suitable for the parenteral, oral, inhalatory and sublingual administrations, while their high biologic activity allows to obtain effective pharmacological effects even with low amounts.
- suitable formulations comprise capsules, tablets, syrups, solutions or sterile injectable lyophils, aerosols.
- the compounds of the invention can be administered in amounts comprised between 0.01 mg/Kg and 10 mg/Kg for the treatment of arthritis, asthma, inflammation, tumoral growth, gastro intestinal hypermotility, Huntington disease, neuritis, neuralgia, migraine, hypertension, incontinence of urine, urticaria, symptoms of the carcinoid syndrome, influenza and cold.
- amino acids are consistent with the recommendations of the IUPAC-IUB Joint Commission on Biochemical nomenclature (Eur. J. Biochem. 1984, 138:9); the amino acids are meant in the L configuration if not otherwise specified.
- NKA neurokinin A
- SP substance P
- Dap 2,3-diaminopropionic acid
- SeCys selenocysteine
- PEG polyethylene glycol
- 1-Nal 1-naphthyl-alanine
- Cha cyclohexylalanine
- gal galactose
- hSer homoserine
- hCys homocysteine
- Boc tert-butyloxycarbonyl
- PAM phenylacetamidomethyl
- Fmoc 9-fluorenylmethyloxycarbonyl
- PyBop benzotriazol-1-yl-oxypyrrolidinephosphonium hexafluorophosphate
- DIEA diisopropylethylamine
- Fm fluorenylmethyl
- DCC dicyclohexylcarbodiimide
- DMF N-N' dimethylformamide
- DCM dichloromethane
- the peptide was assembled in solid phase, through Boc chemistry, on a PAM resin functionalized with Boc-Leu, using a ABI 430A Applied Biosystem synthesizer. The standard deprotection and coupling cycles were used. The following amino acids were coupled in the order with 0.651 g of Boc-Leu-PAM resin (scale 0.5 mmol): Boc-Dap(Fmoc)-OH (1.5 mmol; PyBop 1.5 mmol, DIEA 3 mmol); Boc-Phe-OH, Boc-Trp(CHO)-OH and Boc-Asp(OFm)-OH (2 mmol and DCC/HOBt as the coupling agent).
- the side chains of the two amino acids were deprotected using two deprotection cycles with a solution 20% piperidine in DMF of 3 and 7 min respectively. Repeated washings were then carried out with the following solvents: DCM, DMF, DCM, and i-PrOH.
- the cyclization was carried out adding PyBop (1.5 mmol) and DIEA (3 mmol) to the suspended resin in 15 ml of DMF. The solution was reacted overnight. Afterwards washing cycles were carried out using DMF and DCM.
- Fmoc-Cys( ⁇ -D-Gal(Ac)4)-OH (1.5 mmol) was coupled to the monocyclic peptide using PyBop as coupling agent (0.78 g, 1.5 mmol) and DIEA (3 mmol) as the base. The reaction was carried out overnight.
- the Cys-Fmoc group deprotection was carried out as described for the side chains of Dap and Asp.
- the peptide was assembled in solid phase, through the Boc chemistry according to a procedure identical to the procedure for the cyclo[Cys 1 ( ⁇ -D-Gal)-Asp 2 -Trp 3 -Phe 4 -Dap 5 -Leu 6 )cyclo(2 ⁇ -5 ⁇ ) compound, using Fmoc-Ser 1 ( ⁇ -D-Gal-(Ac) 4 )-OH instead of Fmoc-Cys 1 ( ⁇ -D-Gal(Ac) 4 )-OH.
- the peptide was assembled in solid phase, through the Fmoc chemistry, on a Macrosorb SPR resin functionalized with the p-hydroxymethylbenzoic acid, a base labile linker, using a Milligen 9010 synthesizer. The standard deprotection and coupling cycles were used.
- the esterification of the first residue was carried out with 0.6 mmol (triple excess) of (Fmoc-Leu) 2 O in DMF.
- the coupling of Fmoc-Dap(Boc)-OH, Fmoc-Phe-OH, Fmoc-Trp-OH and Fmoc-Asp(OtBu)-OH was carried out with 0.8 mmol of the amino acid and HOBT/DCC as the activating agent.
- the deprotection of the two amino acidic side chains was carried out with a solution TFA/H 2 O (90:10).
- the cyclization was performed by adding HOBt/DCC (1.6 mmol) to the resin suspended in DMF. The reaction was carried out overnight. After the Asp-amino group deprotection, Fmoc-Asn( ⁇ -D-Gal)-OH (0.8 mmol) was coupled to the peptide using PyBop (0.8 mmol) as the coupling agent and DIEA (1.2 mmol) as the base. The reaction was carried out overnight. The removal of the monocyclic peptide from the resin was carried out by adding 10 ml of a 1 M NaOH solution to 1 g of peptidyl-resin at 4°C. The reaction was effected for 15 min at room temperature.
- the solution was filtered, and the filtrate was collected into a beaker containing 10 ml of an aqueous solution of 10% acetic acid.
- the resin was washed with water and the pH of the filtrate adjusted at 7 with acetic acid.
- 48 mg of PyBop (1 eq.) and 46 ⁇ l of DIEA (3 eq.) were added to 0.1 mmol of the peptide (3) dissolved in 93 ml of DMF (1 mM), keeping the pH between 8 and 8.5.
- the solution was reacted overnight, the progress of the reaction was determined through analytic HPLC.
- the solution was dried and the product purified with preparative HPLC.
- the capability of the products of the present invention of acting as antagonists of the neurokinin A receptor was evaluated using a preparation in which the biologic response produced by tachykinins and by the correlated peptides is determined exclusively by the neurokinin A receptor (NK-2 receptor).
- the deferent conduit which is contracted by the tachykinins in a dose-dependent manner, was used as the experimental model.
- the determination of the peptide activity was carried out using a concentration of ⁇ -Ala 8 -NKA[4-10] (3 nM), as the agonist, which produces a response equal to 45% of the highest response.
Landscapes
- Health & Medical Sciences (AREA)
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Medicinal Chemistry (AREA)
- General Health & Medical Sciences (AREA)
- Life Sciences & Earth Sciences (AREA)
- Chemical Kinetics & Catalysis (AREA)
- General Chemical & Material Sciences (AREA)
- Public Health (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Veterinary Medicine (AREA)
- Pharmacology & Pharmacy (AREA)
- Animal Behavior & Ethology (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Engineering & Computer Science (AREA)
- Molecular Biology (AREA)
- Biochemistry (AREA)
- Proteomics, Peptides & Aminoacids (AREA)
- Genetics & Genomics (AREA)
- Biophysics (AREA)
- Neurosurgery (AREA)
- Biomedical Technology (AREA)
- Neurology (AREA)
- Pulmonology (AREA)
- Pain & Pain Management (AREA)
- Rheumatology (AREA)
- Orthopedic Medicine & Surgery (AREA)
- Dermatology (AREA)
- Psychology (AREA)
- Immunology (AREA)
- Cardiology (AREA)
- Urology & Nephrology (AREA)
- Heart & Thoracic Surgery (AREA)
- Physical Education & Sports Medicine (AREA)
- Nutrition Science (AREA)
- Medicines That Contain Protein Lipid Enzymes And Other Medicines (AREA)
- Peptides Or Proteins (AREA)
- Medicinal Preparation (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Preparation Of Compounds By Using Micro-Organisms (AREA)
Abstract
Description
| Compound | pA2 | Solubility µg/ml |
| Example 1 | 8.4 | 1.800 |
| Comparative example 1 | 6.2 | 2.020 |
| Comparative example 2 | 5.8 | 1.980 |
| MEN 10627 | 8.1 | 15 |
Claims (4)
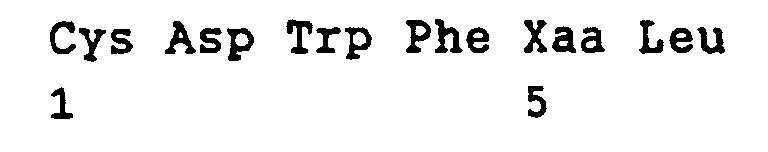
- Compounds of general formula (I) cyclo[X1 1-Z1 2-X2 3-X3 4-Z2 5-X4 6]cyclo(2β-5β) wherein: X1= (D or L)Cys(Y) or (D or L)SeCys(Y), Z1=Asp and Z2=Dap or Z1= Dap and Z2=Asp and X2, X3 and X4 = natural or synthetic hydrophobic amino acids, having Z1, Z2, X2, X3 and X4 the same D or L configuration; Y= is a glycosidic group selected from the aldo or keto hexoses in the furanose or pyranose form, bound to the cysteine through an a or β thioacetalic bond or a cyclithol or a polyvinyl alcohol or PEG, constituted by 5 to 10 monomeric unities, bound to the cysteine with a thioether bond.
- Compounds according to claim 1 wherein: X1= (D or L)Cys(Y) or (D or L)SeCys(Y), Z1=Asp and Z2=Dap or Z1= Dap and Z2=Asp and X2 = Trp or 1-Nal, X3 = Phe and X4 = Leu or Cha, having Z1, Z2, X2, X3, and X4 the same D or L configuration and Y= being defined as in claim 1.
- Pharmaceutical compositions containing a compound of the claims 1 or 2, mixed with a suitable vehicle.
- Use of the compounds of claims 1 or 2 for preparing medicaments for the treatment of arthritis, asthma, inflammation, tumoral growth, gastro-intestinal hypermotility, Huntington disease, neuritis, neuralgia, migraine, hypertension, urine incontinence, urticaria, symptoms of carcinoid syndrome, influenza and cold.
Applications Claiming Priority (3)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| IT96MI000401A IT1283171B1 (en) | 1996-03-01 | 1996-03-01 | SOLUBLE COMPOUNDS ANTAGONISTS OF TACHYCININS THEIR PREPARATION AND THEIR USE IN PHARMACEUTICAL COMPOSITIONS |
| ITMI960401 | 1996-03-01 | ||
| PCT/EP1997/000917 WO1997031941A2 (en) | 1996-03-01 | 1997-02-26 | Soluble tachykinin antagonists, the preparation and use thereof |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| EP0894093A2 EP0894093A2 (en) | 1999-02-03 |
| EP0894093B1 true EP0894093B1 (en) | 2002-05-22 |
Family
ID=11373464
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP97905089A Expired - Lifetime EP0894093B1 (en) | 1996-03-01 | 1997-02-26 | Soluble tachykinin antagonists, the preparation and use thereof |
Country Status (10)
| Country | Link |
|---|---|
| US (1) | US6075006A (en) |
| EP (1) | EP0894093B1 (en) |
| JP (1) | JP2001502656A (en) |
| KR (1) | KR19990087385A (en) |
| AT (1) | ATE217886T1 (en) |
| AU (1) | AU1877197A (en) |
| CA (1) | CA2247802A1 (en) |
| DE (1) | DE69712750D1 (en) |
| IT (1) | IT1283171B1 (en) |
| WO (1) | WO1997031941A2 (en) |
Families Citing this family (4)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| RU2384571C2 (en) | 2004-04-28 | 2010-03-20 | Такеда Фармасьютикал Компани Лимитед | Condensed quinoline derivative and use thereof |
| WO2009072643A1 (en) | 2007-12-03 | 2009-06-11 | Takeda Pharmaceutical Company Limited | Nitrogen-containing heterocyclic compound and use thereof |
| USRE49686E1 (en) | 2008-09-19 | 2023-10-10 | Takeda Pharmaceutical Company Limited | Nitrogen-containing heterocyclic compound and use of same |
| WO2017073764A1 (en) * | 2015-10-30 | 2017-05-04 | 国立大学法人京都大学 | Novel nk3-receptor agonist |
Family Cites Families (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPH06509571A (en) * | 1991-08-08 | 1994-10-27 | エイ・メナリニ・インダストリー・ファーマシューティケ・リウニテ・エスアールエル | Process for producing cyclic hexapeptides as tachykinin antagonists and pharmaceutical compounds thereof |
| IT1262902B (en) * | 1992-04-15 | 1996-07-22 | Menarini Farma Ind | TRICYCLIC COMPOUNDS ANTAGONISTS OF TACHYCHININS, THEIR PREPARATION AND THEIR USE IN PHARMACEUTICAL COMPOSITIONS. |
-
1996
- 1996-03-01 IT IT96MI000401A patent/IT1283171B1/en active IP Right Grant
-
1997
- 1997-02-26 WO PCT/EP1997/000917 patent/WO1997031941A2/en not_active Ceased
- 1997-02-26 AT AT97905089T patent/ATE217886T1/en active
- 1997-02-26 JP JP09530587A patent/JP2001502656A/en active Pending
- 1997-02-26 EP EP97905089A patent/EP0894093B1/en not_active Expired - Lifetime
- 1997-02-26 CA CA002247802A patent/CA2247802A1/en not_active Abandoned
- 1997-02-26 US US09/125,542 patent/US6075006A/en not_active Expired - Fee Related
- 1997-02-26 KR KR1019980706800A patent/KR19990087385A/en not_active Withdrawn
- 1997-02-26 AU AU18771/97A patent/AU1877197A/en not_active Abandoned
- 1997-02-26 DE DE69712750T patent/DE69712750D1/en not_active Expired - Lifetime
Also Published As
| Publication number | Publication date |
|---|---|
| US6075006A (en) | 2000-06-13 |
| DE69712750D1 (en) | 2002-06-27 |
| EP0894093A2 (en) | 1999-02-03 |
| JP2001502656A (en) | 2001-02-27 |
| ITMI960401A0 (en) | 1996-03-01 |
| ATE217886T1 (en) | 2002-06-15 |
| WO1997031941A2 (en) | 1997-09-04 |
| IT1283171B1 (en) | 1998-04-16 |
| WO1997031941A3 (en) | 1997-10-09 |
| CA2247802A1 (en) | 1997-09-04 |
| ITMI960401A1 (en) | 1997-09-01 |
| KR19990087385A (en) | 1999-12-27 |
| AU1877197A (en) | 1997-09-16 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| CN113330024B (en) | Method for preparing GIP/GLP1 dual agonist | |
| AU625598B2 (en) | Peptides having t cell helper activity | |
| US10407468B2 (en) | Methods for synthesizing α4β7 peptide antagonists | |
| Schiller et al. | Synthesis of side‐chain to side‐chain cyclized peptide analogs on solid supports | |
| EP1511761B1 (en) | Process for production of cyclic peptides | |
| KR20170026326A (en) | Method for preparing amg 416 | |
| JP2008534628A (en) | Process for the production of peptide derivatives | |
| CA2325572A1 (en) | Process for the preparation of resin-bound cyclic peptides | |
| US12162957B2 (en) | Process for the preparation of plecanatide | |
| AU664855B2 (en) | Lanthionine bridged peptides | |
| EP3941936A2 (en) | Process of preparation of glucagon-like peptide-1 (glp-1) receptor agonists and their analogs | |
| Schiller et al. | Synthesis and opioid activity profiles of cyclic dynorphin analogs | |
| CN106854235B (en) | A solid-phase fragment method to synthesize carbetocin | |
| US6075006A (en) | Soluble tachykinin antagonists, the preparation and use thereof | |
| Shenbagamurthi et al. | Structure-activity relationships in the dodecapeptide. alpha. factor of Saccharomyces cerevisiae | |
| US5731285A (en) | Tachiquinine antagonist tricyclic compounds, preparation of same and pharmaceutical compositions containing such compounds | |
| US6673769B2 (en) | Lanthionine bridged peptides | |
| KR100405009B1 (en) | Bicyclic tachykinin antagonists, their use in medicaments and their use in pharmaceutical compositions | |
| EP0636146B1 (en) | Tachykinin antagonist tricyclic compounds, preparation of same and pharmaceutical compositions containing such compounds | |
| CN114805480A (en) | Preparation method of octreotide | |
| WO2009150657A1 (en) | Improved process for preparation of eptifibatide by fmoc solid phase synthesis | |
| Thompson et al. | Synthesis of α-aminosuccinimide-containing peptides in Fmoc-based SPPS | |
| Ten Kortenaar et al. | Formation of open-chain asymmetrical cystine peptides on a solid support. Synthesis of pGlu-Asn-Cyt-Pro-Arg-Gly-OH | |
| CN114057829A (en) | Solid-phase synthesis method of N-methylated polypeptide | |
| EP0606222A1 (en) | Cyclic hexapeptides as tachyquinin antagonists, their preparation and pharmaceutical compositions thereof |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| 17P | Request for examination filed |
Effective date: 19980921 |
|
| AK | Designated contracting states |
Kind code of ref document: A2 Designated state(s): AT BE CH DE DK ES FI FR GB GR IE IT LI LU MC NL PT SE |
|
| GRAG | Despatch of communication of intention to grant |
Free format text: ORIGINAL CODE: EPIDOS AGRA |
|
| 17Q | First examination report despatched |
Effective date: 20010530 |
|
| GRAG | Despatch of communication of intention to grant |
Free format text: ORIGINAL CODE: EPIDOS AGRA |
|
| GRAH | Despatch of communication of intention to grant a patent |
Free format text: ORIGINAL CODE: EPIDOS IGRA |
|
| GRAH | Despatch of communication of intention to grant a patent |
Free format text: ORIGINAL CODE: EPIDOS IGRA |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: NL Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: LI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: IT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT;WARNING: LAPSES OF ITALIAN PATENTS WITH EFFECTIVE DATE BEFORE 2007 MAY HAVE OCCURRED AT ANY TIME BEFORE 2007. THE CORRECT EFFECTIVE DATE MAY BE DIFFERENT FROM THE ONE RECORDED. Effective date: 20020522 Ref country code: GR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: FR Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: FI Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: CH Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: BE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 Ref country code: AT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020522 |
|
| REF | Corresponds to: |
Ref document number: 217886 Country of ref document: AT Date of ref document: 20020615 Kind code of ref document: T |
|
| REG | Reference to a national code |
Ref country code: GB Ref legal event code: FG4D |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: EP |
|
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: FG4D |
|
| REF | Corresponds to: |
Ref document number: 69712750 Country of ref document: DE Date of ref document: 20020627 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020822 Ref country code: PT Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020822 Ref country code: DK Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020822 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20020823 |
|
| NLV1 | Nl: lapsed or annulled due to failure to fulfill the requirements of art. 29p and 29m of the patents act | ||
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: ES Free format text: LAPSE BECAUSE OF FAILURE TO SUBMIT A TRANSLATION OF THE DESCRIPTION OR TO PAY THE FEE WITHIN THE PRESCRIBED TIME-LIMIT Effective date: 20021128 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| EN | Fr: translation not filed | ||
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LU Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20030226 Ref country code: IE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20030226 Ref country code: GB Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20030226 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: MC Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20030228 |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| 26N | No opposition filed |
Effective date: 20030225 |
|
| GBPC | Gb: european patent ceased through non-payment of renewal fee | ||
| REG | Reference to a national code |
Ref country code: IE Ref legal event code: MM4A |