EP0495485B1 - Phenoxyalkylcarboxylic acid derivatives, process of preparing the same and anti-allergic agents - Google Patents
Phenoxyalkylcarboxylic acid derivatives, process of preparing the same and anti-allergic agents Download PDFInfo
- Publication number
- EP0495485B1 EP0495485B1 EP92100614A EP92100614A EP0495485B1 EP 0495485 B1 EP0495485 B1 EP 0495485B1 EP 92100614 A EP92100614 A EP 92100614A EP 92100614 A EP92100614 A EP 92100614A EP 0495485 B1 EP0495485 B1 EP 0495485B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- general formula
- denotes
- compound
- group
- hydrogen atom
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Expired - Lifetime
Links
- 0 CCSc1c(*)c(O)c(*)cc1 Chemical compound CCSc1c(*)c(O)c(*)cc1 0.000 description 11
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C323/00—Thiols, sulfides, hydropolysulfides or polysulfides substituted by halogen, oxygen or nitrogen atoms, or by sulfur atoms not being part of thio groups
- C07C323/22—Thiols, sulfides, hydropolysulfides or polysulfides substituted by halogen, oxygen or nitrogen atoms, or by sulfur atoms not being part of thio groups containing thio groups and doubly-bound oxygen atoms bound to the same carbon skeleton
Definitions
- the present invention concerns novel phenoxyalkylcarboxylic acid derivatives having a strong and selective leukotriene antagonistic action and useful for prophylaxis and therapy of allergic diseases as asthma, and processes of preparing the same.
- leukotrienes which are metabolites of arachidonic acid through 5-lipoxygenase pathway, are the components of SRS-A (slow reactiong substance of anaphylaxis) considered to be a major etiogenic substance of immediate type allergic diseases such as bronchial asthma and so on.
- leukotriene antagonists are expected as a useful anti-allergic agent.
- phenoxyalkylcarboxylic acid derivatives act as leukotriene antagonists (Japanese Laid-open Publication No. Hei 2-1459 corresponding to EP 0 332 109 and US 4,985,585), but there has been an increasing need for the preparation of compounds having activity in vivo.
- phenoxyalkylcarboxylic acid derivarives represented by the following general formula (1) have a strong and selective leukotriene antagonistic action, and completed the present invention.
- R1 denotes a hydrogen atom, methyl group and ethyl group.
- the compound of the general formula (1) can be prepared through the routes mentioned below.
- the compound of the present invention showed strong antagonistic action to leukotriene D4 in the isolated guinea pig trachea. Furthermore, the compound intravenously injected inhibited leukotriene D4 -induced bronchoconstriction even at a lower dose level.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Acyclic And Carbocyclic Compounds In Medicinal Compositions (AREA)
Description
- The present invention concerns novel phenoxyalkylcarboxylic acid derivatives having a strong and selective leukotriene antagonistic action and useful for prophylaxis and therapy of allergic diseases as asthma, and processes of preparing the same.
- The leukotrienes (leukotriene C₄, D₄, E₄), which are metabolites of arachidonic acid through 5-lipoxygenase pathway, are the components of SRS-A (slow reactiong substance of anaphylaxis) considered to be a major etiogenic substance of immediate type allergic diseases such as bronchial asthma and so on.
- Hence, leukotriene antagonists are expected as a useful anti-allergic agent.
- The inventors of the present invention had formerly found that phenoxyalkylcarboxylic acid derivatives act as leukotriene antagonists (Japanese Laid-open Publication No. Hei 2-1459 corresponding to EP 0 332 109 and US 4,985,585), but there has been an increasing need for the preparation of compounds having activity in vivo.
- As the result of diligent study concerning leukotriene antagonists, the inventors of the present invention have found that phenoxyalkylcarboxylic acid derivarives represented by the following general formula (1) have a strong and selective leukotriene antagonistic action, and completed the present invention.

wherein R¹ denotes a hydrogen atom, methyl group and ethyl group. - According to the present invention, the compound of the general formula (1) can be prepared through the routes mentioned below.
- (1) A compound of the following general formula (1a) can be prepared by allowing a compound of the general formula (2) to react with a compound of the general formula (3)


The reaction is preferably conducted in an organic solvent, for example, acetone, methylethylketone, diethylketone or dimethylformamide at a reaction temperature of room temperature to solvent refluxing temperature. Then, the presence of an inorganic base, for example, potassium carbonate or sodium carbonate and further the addition of potassium iodide are also favorable.
A compound of the general formula (1a) can be converted into the corresponding carboxylic acid compound by a conventional method. - (2) A compound represented by the general formula (1) can be prepared by allowing a compound of the general formula (4) to react with a compound of the formula (5), if necessary, followed by hydrolysis thereof.


- (3) A compound represented by the general formula (7) can be prepared by allowing a compound of the general formula (6) to undergo a catalytic hydrogenation with hydrogen or to react with a reducing agent.


The catalytic hydrogenation with hydrogen is preferably conducted, under the ordinary pressure or an increased pressure, in methanol, ethanol, tetrahydrofuran or dimethylformamide at a reaction temperature of 0 °C to the solvent refluxing temperature. The catalyst to be used is favorably a heterogeneous or homogeneous palladium, nickel, rhodium or ruthenium and furthermore the use of asymmetric catalyst is also favorable.
In case of the reaction with reducing agent, it is preferably allowed to react with the reducing agent, for example, sodium borohydride or lithium aluminum hydride in methanol, ethanol, tetrahydrofuran or dimethylformamide at a reaction temperature of cooling with ice to the solvent refluxing temperature.
In case R¹ denotes methyl group or ethyl group in the general formula (7), it can be converted in the corresponding carboxylic acid compound by a conventional method.
Furthermore, the compound represented by the general formula (1) has an asymmetric carbon on 1-position of 1-hydroxyethyl group and so two kinds of optical isomer exist basing on the asymmetric carbon, but the respective isomers or the mixture thereof are all involved in the present invention.
The two kinds of optical isomer can be optically resolved, for example, by forming diastereomeric salt of base such as (S)-(-)-1-(1-naphthyl) ethylamine with the corresponding carboxylic acid compound or, by taking out with separation using an optically active column. This optical resolution is possible in the compound of the general formula (1) as well as in that of the general formula (7).
Further, the compound represented by the general formula (1), if desired, can be converted into the salt thereof by a conventional method. As the salt thereof, the salt of sodium, potassium, calcium or aluminum is exemplified. - In the following, the present invention is illustrated by the concrete examples.
-
(i) A mixture of 8.7g of ethyl 4-(3-hydroxy-6-(1-hydroxyethyl)-2-propylphenoxy) lactate, 10.21g of (4-(3-bromopropylthio)-2-hydroxy-3-propylphenyl) ethanone, and 7.79g of potassium carbonate and acetone (70ml) was refluxed with heating and stirring. To the reaction mixture were added 2.79 g of (4-(3-bromopropylthio)-2-hydroxy-3-propylphenyl) ethanone after 9 hours and 3.87 g of pottasium carbonate after 9 hours, 11 hours and 16 hours respectively. Then, after the mixture was refluxed with heating and stirring for 18 hours in all, the inorganic matter was filtered off and the filtrate was distilled off under the reduced pressure,
The residue was purified through medium pressure silica gel column chromatography (benzene: ethyl acetate = 9:1) to afford 13.6 g of crude ethyl 4-[3-[3-(4-acetyl-3-hydroxy-2-propylphenylthio)propoxy]-6-(1-hydroxyethyl)-2-propylphenoxy]butanoate as an oily product (86.5 %).
(ii) To a solution of 6.60 g of ethyl 4-[3-[3-(4-acetyl-3-hydroxy-2-propylphenylthio)propoxy]-6-(1-hydroxyethyl)-2-propylphenoxy] butanoate in 10 ml of ethanol was added a solution of 1.41 g of sodium hydroxide in 10 ml of water. After stirred at room temperature for 5 minutes, the mixture was cooled by addition of ice water and washed with ether. To the aqueous layer was acidified with hydrochloric acid and was extracted with ethyl acetate. The organic layer was washed with water, dried over sodium sulfate and concentrated. After the residue was purified through medium pressure silica gel column chromatography (normal phase with methylene chloride: ethanol = 100:3, then reverse phase with methanol: water = 19:1), it was recrystallized from methanol-water to afford 1.98 g of the aimed product as colorless crystal (31.6 %).
Melting point 85 - 86 °CElementary analysis (%) for C₂₉H₄₀O₇S Calculated value (Observed value) C: 65.39 (65.20) H: 7.57 (7.63) -
- 16.4 g of ethyl 4-(6-acetyl-3-hydroxy-2-propylphenoxy) lactate was dissolved into 90 ml of ethanol and added with 1.2 g of 5 % palladium-on-charcoal, the mixture of which was then subjected to catalytic hydrogenation with hydrogen under the atmospheric pressure with water cooling.
- After completion of the reaction, the catalyst was filtered off and the filtrate was concentrated. The residue was purified through medium pressure silica gel column chromatography (benzene: ethyl acetate = 7:3) to afford 14.0 g of the aimed product being light yellow oily (84.8 %).
¹H-NMR (CDCl₃) δ:
0.98 (3H, t, J = 7Hz),
1.28 (3H, t, J = 7Hz),
1.48 (3H, d, J = 6Hz),
1.58 (2H, m),
2.14 (2H, m),
2.4 - 2.6 (5H, m),
3.8 - 3.9 (2H, m),
4.17 (2H, q),
- 5.1 (1H, m),
5.65 (1H, s),
6.56 (1H, d, J = 9Hz),
7.11 (1H, d, J = 9Hz). -
- To a solution of 15.0 g of ethyl 4-[6-acetyl-3-(3-chloropropoxy)-2-propylphenoxy] butanoate in 195 ml of methanol was added 2.9 g of sodium borohydride in portions of small quantity under ice cooling and the mixture was stirred at the same temperature for two hours. To the reaction mixture was added 100 ml of water and the solvent was distilled off under reduced pressure. Then, the mixture was acidified with 2N-hydrochloric acid under ice cooling, followed by extraction with ethyl acetate. The organic layer was distilled off under reduced pressure to afford 14.6 g of the aimed product as light yellow oily matter (96.8 %).
NMR (CDCl₃) δ:
0.96 (3H, t, J = 7Hz),
1.27 (3H, t, J = 7Hz),
1.40 - 1.80 (2H, m),
1.47 (3H, d, J = 6.6Hz),
2.00 - 2.40 (4H, m),
2.40 - 2.70 (5H, m),
3.70 - 4.00 (4H, m),
4.00 - 4.30 (4H, m),
5.10 (1H, q, J = 6.6Hz),
6.70 (1H, d, J = 8.4Hz),
7.20 (1H, d, J = 8.8Hz). - To a solution of 14.6 g of ethyl 4-(3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] butanoate in 85 ml of ethanol was dropwise added 33.3 ml of an aqueous solution of 1.67 g of sodium hydroxide under ice cooling. After stirred for an hour as it was and then allowed to react for 4 hours at room temperature, 5 ml of aqueous solution of 250 mg of sodium hydroxide was twice supplemented to the reaction mixture with interval of an hour to allow the reaction proceeds. After addition of 1N hydrochloric acid under ice cooling to acidify and distillation off of the solvent under reduced pressure, the reaction mixture was alkalilized with 5 % aqueous solution of sodium hydroxide and washed with ethyl acetate. The aqueous layer was acidified with 1N hydrochloric acid and extracted with ethyl acetate. The organic layer was dried over anhydrous sodium sulfate after washed with water and saturated aqueous solution of sodium chloride. It was subjected to distillation off of the solvent under reduced pressure to afford 12.9 g of the aimed product as light yellow oily matter (yield 95.1 %).
NMR (CDCl₃) δ:
0.95 (3H, t, J = 7.3Hz),
1.38 - 1.80 (2H, m),
1.47 (3H, d, J = 6.6Hz),
2.00 - 2.40 (4H, m),
2.40 - 2.75 (4H, m),
3.62 - 3.94 (4H, m),
4.08 (2H, t, J = 5.7Hz),
5.12 (1H, q, J = 6.6Hz),
6.66 (1H, d, J = 8.8Hz),
7.22 (1H, d, J = 8.8Hz). - To a solution of 5.0 g of 4-[3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] butanoic acid in 7 ml of ethyl acetate was added 2.15 ml of (s)-(-)-1-(1-naphthyl) ethylamine, which was then allowed to stand for overnight. The precipitate was collected by filtration and dried after washing with cold ethyl acetate to afford 1.07 g of crude salt. The crude salt was three times recrystallized from ethyl acetate to afford 68.0 mg of (s)-(-)-1-(1-naphthyl) ethylamine salt of (+)-4-[3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] butanoic acid (melting point 124.0 - 125.0 °C, [α]


- The crude salt was three times recrystallized with ethyl acetate to afford 141.4 mg as colorless needle crystal of (R)-(+)-1-(1-naphthyl) ethylamine salt of (-)-4-[3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] lactic acid (melting point 123.0 - 124.5 °C [α]


- A mixed solution of 55.9 mg of (-)-4-[3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] butanoic acid, 39.3 mg of (2-hydroxy-4-mercapto-3-propylphenyl) ethanone, 51.7 mg of potassium carbonate and 1 ml of dimethylformamide was stirred at room temperature for 6 hours. The reaction mixture was poured into ice water, acidified with 1N-hydrochloric acid, and extracted with ethyl acetate. After washed with water and saturated aqueous solution of sodium chloride, the organic layer was dried over anhydrous sodium sulfate and subjected to distillation off of the solvent under reduced pressure to afford residue, which was then purified through silica gel column chromatography (methylene chloride: methanol = 20:1) and further purified through preparative thin layer chromatography (methylene chloride: methanol = 15:1) to afford 29.8 mg of yellow oily matter as the title compound (35.9 %). [α]

NMR (CDCl₃) δ:
0.84 - 1.17 (6H, m),
1.37 - 1.75 (4H, m),
1.49 (3H, d, J = 6.6Hz),
2.09 - 2.36 (4H, m),
2.40 - 2.78 (6H, m),
2.58 (3H, s),
3.19 (2H, t, J = 7.5Hz),
3.74 - 4.13 (6H, m),
5.12 (1H, q, J = 6.2Hz),
6.64 (1H, d, J = 8.8Hz),
6.76 (1H, d, J = 8.4Hz),
7.23 (1H, d, J = 8.4Hz),
7.50 (1H, d, J = 8.4Hz),
12.73 (1H, s). - A mixed solution of 42.6 mg of (+)-4-[3-(3-chloropropoxy)-6-(1-hydroxyethyl)-2-propylphenoxy] butanoic acid, 30.0 mg of (2-hydroxy-4-mercapto-3-propylphenyl) ethanone, 39.4 mg of potassium carbonate and 1 ml of dimethylformamide was stirred at room temperature for 4 hours. The reaction mixture was poured into ice water, acidified with 1N hydrochloric acid and extracted with ethyl acetate. After washed with water and saturated aqueous solution of sodium chloride, the organic layer was dried over anhydrous sodium sulfate and subjected to distillation off of the solvent under reduced pressure to afford residue. The residue was purified through silica gel column chromatography (methylene chloride: ethanol = 20:1) and further through preparative thin layer chromatography (methylene chloride: methanol = 1.5:1) to afford 16.1 mg of yellow oily water as the title compound (yield 25.4 %). [α]

NMR (CDCl₃) δ:
0.84 - 1.17 (6H, m),
1.37 - 1.75 (4H, m),
1.48 (3H, d, J = 6.6Hz),
2.09 - 2.36 (4H, m),
2.40 - 2.78 (6H, m),
2.58 (3H, S),
3.19 (2H, t, J = 7.5Hz),
3.56 (2H, brs),
3.81 - 4.07 (4H, m),
5.12 (1H, q, J = 6.6Hz),
6.64 (1H, d, J = 8.8Hz),
6.76 (1H, d, J = 8.8Hz),
7.23 (1H, d, J = 8.4Hz),
7.50 (1H, d, J = 8.4Hz),
12.74 (1H, S). - Male Hartly guinea pigs weighing about 450 g were anesthetized with sodium pentobarbital (30 mg/kg,i.p.) and the change in intra-tracheal pressure was measured according to modified Konzett-Rössler method (J.Harvey, et al., J.Phamacol. Method, 9, 147-155, 1983). Bronchoconstriction was indused by i.v. injection of leukotriene D₄ (3»g/kg) into the left external jugular vein. Animals were pretreated with i.v. injection of indomethacin and propranolol. The compound of Example 1 as sodium salt solution was i.v. administered 3 min before leukotriene D₄ injection. The result was showed in Table 1.
- The compound of the present invention showed strong antagonistic action to leukotriene D₄ in the isolated guinea pig trachea. Furthermore, the compound intravenously injected inhibited leukotriene D₄ -induced bronchoconstriction even at a lower dose level.
- These results indicate that the compound in the present invention represented by the general formula (1) are useful for treatment of the diseases caused by leukotrienes, such as bronchial asthma, allergy of eye, nose and gastrointestinal tract, allergic dermatitis, circulatory and so on.
Table 1 Tested Compound Dose (mg/kg, i. v.) Inhibition (%) Example 1 0.125 -0.5 0.25 45.6 0.5 56.8 1.0 71.1
Claims (5)
- A method of preparing a phenoxyalkylcarboxylic acid derivative or an alkali salt thereof represented by the general formula (1)

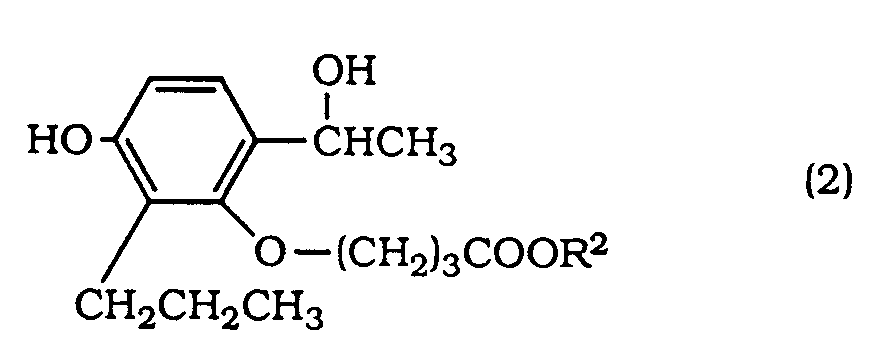


- A method of preparing a compound of the general formula (1)


- A method of preparing a compound of the general formula (7)

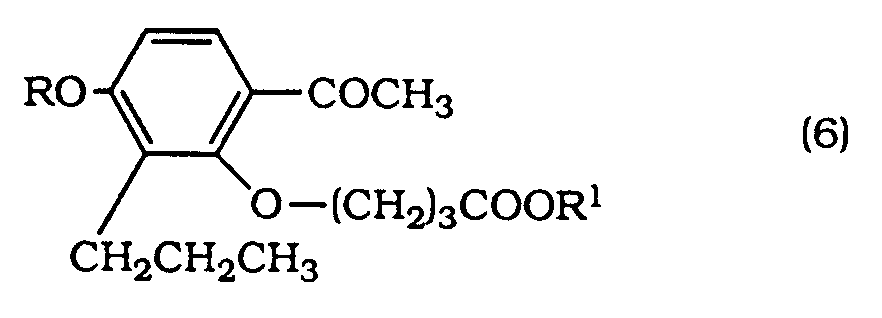
Applications Claiming Priority (4)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP54207/91 | 1991-01-18 | ||
| JP5420791 | 1991-01-18 | ||
| JP3359460A JPH0551359A (en) | 1991-01-18 | 1991-12-26 | Phenyloxyalkylcarboxylic acid derivative and process for producing the same |
| JP359460/91 | 1991-12-26 |
Publications (3)
| Publication Number | Publication Date |
|---|---|
| EP0495485A2 EP0495485A2 (en) | 1992-07-22 |
| EP0495485A3 EP0495485A3 (en) | 1992-08-26 |
| EP0495485B1 true EP0495485B1 (en) | 1995-04-12 |
Family
ID=26394955
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP92100614A Expired - Lifetime EP0495485B1 (en) | 1991-01-18 | 1992-01-15 | Phenoxyalkylcarboxylic acid derivatives, process of preparing the same and anti-allergic agents |
Country Status (8)
| Country | Link |
|---|---|
| US (1) | US5290812A (en) |
| EP (1) | EP0495485B1 (en) |
| CN (1) | CN1063487A (en) |
| AU (1) | AU638626B2 (en) |
| CA (1) | CA2058888C (en) |
| DE (1) | DE69201964T2 (en) |
| ES (1) | ES2073189T3 (en) |
| HU (1) | HU214714B (en) |
Families Citing this family (9)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US4985585A (en) | 1988-03-07 | 1991-01-15 | Kyorin Pharmaceutical Co., Ltd. | Phenoxyalkylcarboxylic acid derivatives and process for their preparations |
| JP2007534771A (en) * | 2004-04-27 | 2007-11-29 | メディシノバ,インコーポレーテッド | Phenoxyalkylcarboxylic acid derivatives in the treatment of inflammatory diseases |
| JP2013119550A (en) | 2011-12-08 | 2013-06-17 | Medicinova Inc | Method for treating nonalcoholic fatty liver disease and nonalcoholic steatohepatitis |
| US8962687B2 (en) | 2012-12-05 | 2015-02-24 | Medicinova, Inc. | Method of treating liver disorders |
| WO2015013395A1 (en) | 2013-07-25 | 2015-01-29 | Medicinova, Inc. | Methods for reducing triglyceride, total cholesterol and low density lipoprotein blood levels |
| US9346754B2 (en) | 2014-05-08 | 2016-05-24 | Medicinova, Inc. | Method of treating advanced non-alcoholic steatohepatitis |
| US20150321989A1 (en) | 2014-05-08 | 2015-11-12 | Medicinova, Inc. | Method of treating idiopathic pulmonary fibrosis |
| CA2950909C (en) | 2014-06-02 | 2023-03-28 | Medicinova, Inc. | Method of inhibiting or treating fibrosis |
| CN117320701A (en) | 2021-05-28 | 2023-12-29 | 美迪诺亚公司 | A method of reducing triglyceride synthesis in the liver |
Family Cites Families (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US4565882A (en) * | 1984-01-06 | 1986-01-21 | G. D. Searle & Co. | Substituted dihydrobenzopyran-2-carboxylates |
| JPH0819041B2 (en) * | 1987-09-10 | 1996-02-28 | 杏林製薬株式会社 | Phenoxyalkylcarboxylic acid derivative and method for producing the same |
| US4985585A (en) * | 1988-03-07 | 1991-01-15 | Kyorin Pharmaceutical Co., Ltd. | Phenoxyalkylcarboxylic acid derivatives and process for their preparations |
-
1991
- 1991-12-30 US US07/814,477 patent/US5290812A/en not_active Expired - Lifetime
-
1992
- 1992-01-02 HU HU9200008A patent/HU214714B/en not_active IP Right Cessation
- 1992-01-07 CA CA002058888A patent/CA2058888C/en not_active Expired - Fee Related
- 1992-01-15 EP EP92100614A patent/EP0495485B1/en not_active Expired - Lifetime
- 1992-01-15 DE DE69201964T patent/DE69201964T2/en not_active Expired - Fee Related
- 1992-01-15 ES ES92100614T patent/ES2073189T3/en not_active Expired - Lifetime
- 1992-01-16 CN CN92100324A patent/CN1063487A/en active Pending
- 1992-01-16 AU AU10288/92A patent/AU638626B2/en not_active Expired
Also Published As
| Publication number | Publication date |
|---|---|
| CA2058888A1 (en) | 1992-07-19 |
| EP0495485A3 (en) | 1992-08-26 |
| AU638626B2 (en) | 1993-07-01 |
| CN1063487A (en) | 1992-08-12 |
| EP0495485A2 (en) | 1992-07-22 |
| DE69201964T2 (en) | 1995-12-14 |
| HUT61979A (en) | 1993-03-29 |
| ES2073189T3 (en) | 1995-08-01 |
| DE69201964D1 (en) | 1995-05-18 |
| CA2058888C (en) | 2003-06-17 |
| AU1028892A (en) | 1992-07-23 |
| HU214714B (en) | 1998-08-28 |
| US5290812A (en) | 1994-03-01 |
| HU9200008D0 (en) | 1992-03-30 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| DE2814556C2 (en) | Substituted phenylacetic acid derivatives, processes for their preparation and pharmaceutical compositions containing them | |
| US5290812A (en) | Phenoxyalkylcarboxylic acid derivatives and process of preparing the same | |
| CZ282400B6 (en) | Benzopyran compounds, process of their preparation, process for preparing intermediates, pharmaceutical preparation and use of such compounds | |
| US4304931A (en) | Biphenylhydroxypropionic acid derivatives | |
| US4188491A (en) | Preparation of therapeutic compounds | |
| KR950014597B1 (en) | Phenoxyalkylcarboxylic acid derivatives and preparation method thereof | |
| US5489592A (en) | 3,4-dihydro-4-oxo-3-(2-propenyl)-1-phthalazineacetic acids and derivatives, their preparations and medicines containing them | |
| McKENNIS JR et al. | meso and dl-2, 3-Diaminosuccinic Acids1 | |
| EP0916657B1 (en) | 1-phenylpyrrolidone derivatives having optical activity | |
| EP0071500A1 (en) | Process for the preparation of 4-aminobutyramide | |
| CN116178190B (en) | A preparation method of proparacaine hydrochloride | |
| JP2000119221A (en) | Method for producing (2,4,5-trifluoro-3-methoxybenzoyl) acetic acid ester derivative and intermediate for producing the same | |
| LU82115A1 (en) | NOVEL ANTI-HYPERTENSIVE QUINAZOLINES, PROCESS FOR THEIR PRODUCTION AND PHARMACEUTICAL COMPOSITION CONTAINING THEM | |
| EP0036030B1 (en) | Process for preparing biotin | |
| JP4449211B2 (en) | 6- (1-fluoroethyl) -5-iodo-4-pyrimidone and process for producing the same | |
| FR2569194A1 (en) | PROCESS FOR THE PREPARATION OF AMINOLACTONE | |
| JPH01163154A (en) | Production of tetrahydrophthalimide based compound, intermediate thereof and production of said intermediate | |
| GB2239242A (en) | Cyclohexanol derivative | |
| SU545250A3 (en) | Method for preparing substituted biphenylyl butyric acid or its esters or its salts | |
| JPS60209569A (en) | N-bicycloheptyl- and n-bicycloheptenyl-imidazole and manufacture | |
| JPH06100526A (en) | Phenoxyalkylcarboxylic acid derivative | |
| JPH06755B2 (en) | (+)-Lattein and method for producing the same | |
| JPH0129176B2 (en) | ||
| CH406190A (en) | Process for preparing 10-hydroxy-2-decene trans oic acid | |
| JPH05194336A (en) | Aminoacrylic acid derivative |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| PUAL | Search report despatched |
Free format text: ORIGINAL CODE: 0009013 |
|
| AK | Designated contracting states |
Kind code of ref document: A2 Designated state(s): BE CH DE ES FR GB IT LI NL SE |
|
| AK | Designated contracting states |
Kind code of ref document: A3 Designated state(s): BE CH DE ES FR GB IT LI NL SE |
|
| 17P | Request for examination filed |
Effective date: 19921230 |
|
| 17Q | First examination report despatched |
Effective date: 19940422 |
|
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| AK | Designated contracting states |
Kind code of ref document: B1 Designated state(s): BE CH DE ES FR GB IT LI NL SE |
|
| REF | Corresponds to: |
Ref document number: 69201964 Country of ref document: DE Date of ref document: 19950518 |
|
| ET | Fr: translation filed | ||
| ITF | It: translation for a ep patent filed | ||
| REG | Reference to a national code |
Ref country code: ES Ref legal event code: FG2A Ref document number: 2073189 Country of ref document: ES Kind code of ref document: T3 |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |
|
| 26N | No opposition filed | ||
| REG | Reference to a national code |
Ref country code: GB Ref legal event code: IF02 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: ES Payment date: 20030103 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: SE Payment date: 20030113 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: GB Payment date: 20030115 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: FR Payment date: 20030117 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: BE Payment date: 20030122 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: CH Payment date: 20030123 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 20030130 Year of fee payment: 12 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: NL Payment date: 20030131 Year of fee payment: 12 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GB Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040115 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040116 Ref country code: ES Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040116 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: LI Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040131 Ref country code: CH Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040131 Ref country code: BE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040131 |
|
| BERE | Be: lapsed |
Owner name: *KYORIN PHARMACEUTICAL CO. LTD Effective date: 20040131 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: NL Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040801 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040803 |
|
| EUG | Se: european patent has lapsed | ||
| GBPC | Gb: european patent ceased through non-payment of renewal fee |
Effective date: 20040115 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: FR Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20040930 |
|
| NLV4 | Nl: lapsed or anulled due to non-payment of the annual fee |
Effective date: 20040801 |
|
| REG | Reference to a national code |
Ref country code: FR Ref legal event code: ST |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: IT Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 20050115 |
|
| REG | Reference to a national code |
Ref country code: ES Ref legal event code: FD2A Effective date: 20040116 |